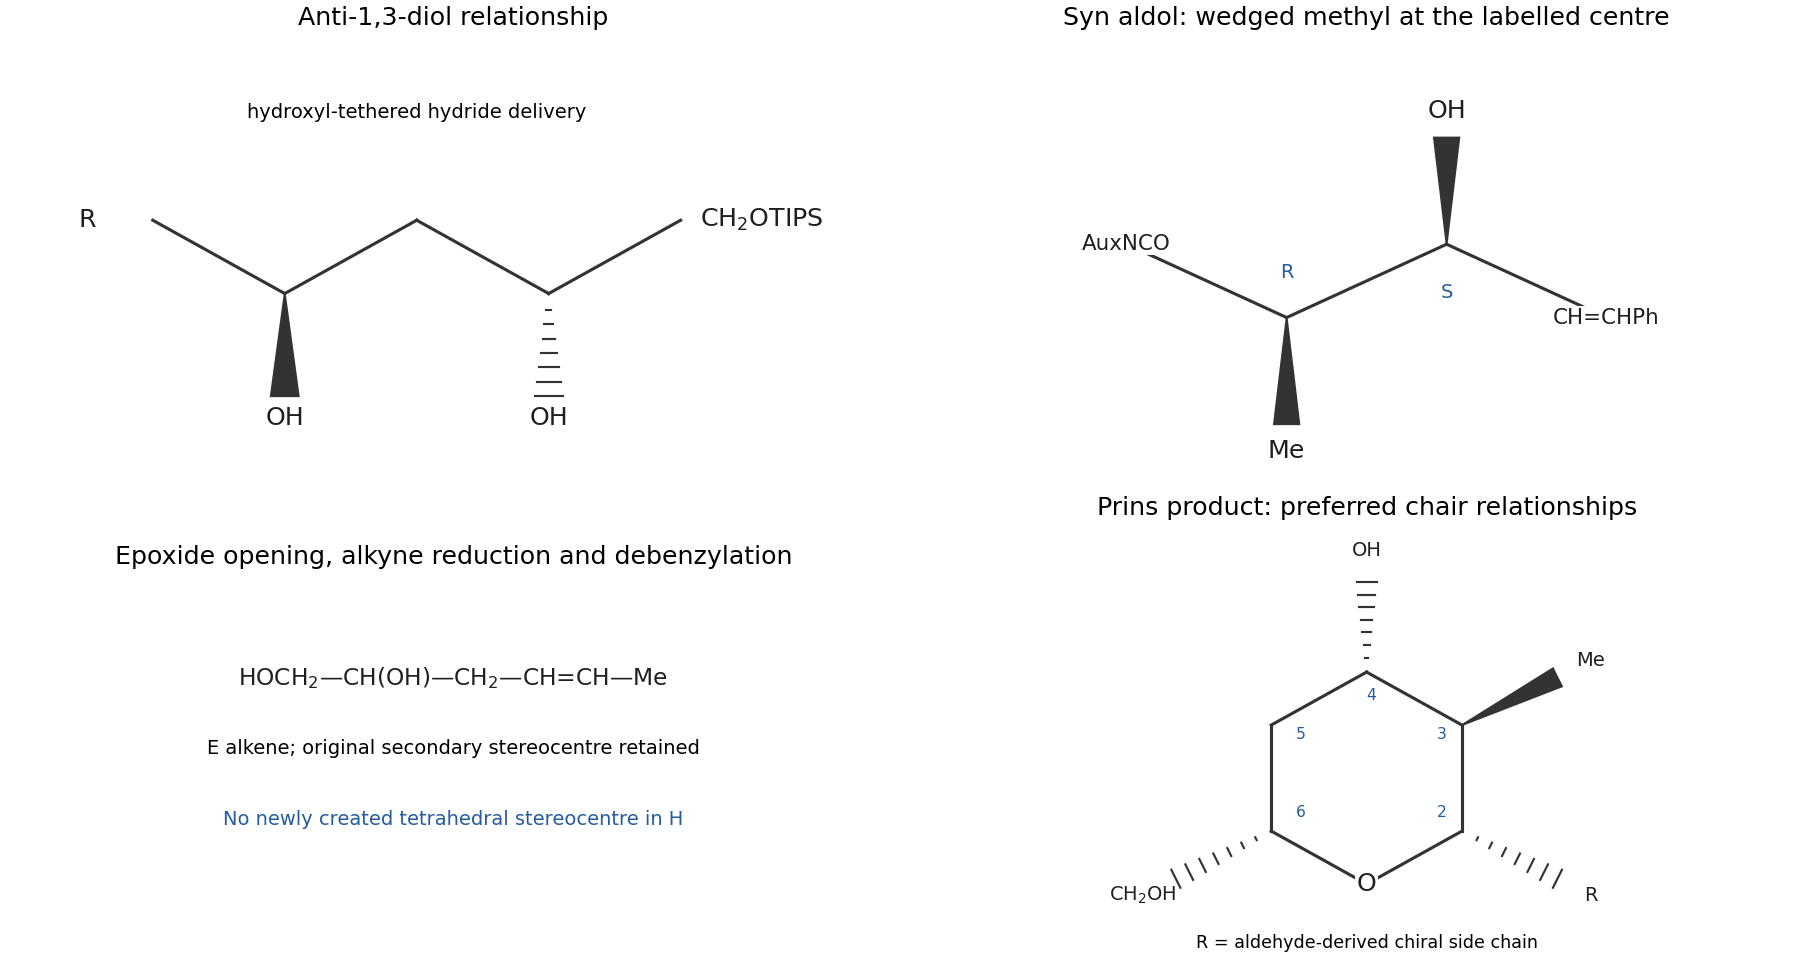
A hydroxyl-directed Evans–Saksena reduction gives the required anti-1,3-diol. A suitable reagent is tetramethylammonium triacetoxyborohydride in an acidic medium, for example acetic acid/acetonitrile. Ligand exchange between the pre-existing alcohol and an acetoxyborohydride group tethers the reducing reagent to the substrate. Intramolecular hydride delivery to the ketone then proceeds through a six-membered arrangement, converting C=O to an alkoxide while conserving all existing stereocentres. Hydrolysis releases the second hydroxyl group.
The preferred directed transition-state conformation minimizes crowding by the larger carbon-chain substituents. It delivers the new C–H bond on the face that leaves the two hydroxyl groups anti, as in B: with the original hydroxyl drawn as a wedge in the printed zigzag, the new hydroxyl is dashed. The intervening methyl stereocentre is retained. Choosing an undirected borohydride reduction would not by itself rationalize this strong substrate-controlled relationship.
Dibutylboron triflate coordinates the imide carbonyl and triethylamine removes the propionyl proton, giving predominantly the Z dibutylboron enolate. Its connectivity is ; the imide auxiliary remains attached, rather than being displaced during enolization. Coordination of cinnamaldehyde's oxygen to boron organizes a six-membered Zimmerman–Traxler transition state.
A chair with the aldehyde's styryl group pseudoequatorial avoids the auxiliary's bulky benzyl side. Formation of the C–C bond and transfer of the aldehyde oxygen into the boron coordination sphere give the boron aldolate, which hydrolyses to D. A Z enolate gives the syn aldol relationship. In the printed product zigzag, the labelled -methyl bond should therefore be a solid wedge, on the same stereochemical face as the neighbouring solid-wedge hydroxyl bond. With the stated connectivity, the labelled carbon is R: priority is imide carbonyl, then , then methyl, then hydrogen. The adjacent hydroxyl centre is S. These assignments refer to the actual auxiliary and product orientation, not an arbitrary choice of an otherwise equivalent chair.
The free allylic hydroxyl can hydrogen-bond to the peracid and accelerate directed oxygen transfer to the nearby double bond. The remote styryl double bond lacks this directing contact. Epoxidation of the hydroxyl-adjacent alkene is therefore favoured, even though both alkenes are in principle oxidizable. Peracid epoxidation is a concerted oxygen transfer: alkene donation to terminal peroxide oxygen, O–O cleavage, proton transfer and carbonyl re-formation occur through a cyclic transition-state arrangement. The alkene's E geometry is retained as a trans relationship between its substituents in the epoxide.
The two alkene faces are diastereotopic because the molecule already contains stereocentres. A plausible substrate-controlled conformation minimizes allylic strain and presents the face opposite the bulky OTBS-bearing part of the chain to the oxidant. Predict epoxide oxygen on the face opposite OTBS, retaining the pre-existing methyl and OTBS configurations. Hydrogen-bond direction, steric approach and conformational population compete; one cannot deduce an exact diastereomeric ratio from the flat drawing alone. The same facial preference has been discussed for this synthetic series.[ref]
Recent results toward the stereoselective synthesis of biologically active natural products by Luiz C. Dias and others. 2007, synthesis section and Scheme 2, epoxidation opposite OTBS
The acetylide attacks the less substituted terminal epoxide carbon by SN2 ring opening. The C–O bond at that carbon breaks, leaving the oxygen attached to the original substituted stereocentre. Protonation gives . Dissolving-metal reduction then gives the E alkene and also cleaves the benzyl ether.
Thus H is with an E double bond, and its original epoxide-derived stereocentre is retained geometrically. No new tetrahedral stereocentre is formed: the attacked epoxide carbon becomes CH. The new stereochemical information is the E alkene geometry. Calling its secondary hydroxyl carbon “newly generated” would confuse changing its functional environment with creating a new stereocentre.
The secondary alcohol of H condenses with the aldehyde under acid catalysis to form a hemiacetal, then loses water to an oxocarbenium ion. Its tethered E alkene attacks the aldehyde-derived carbon through a chair-like six-membered arrangement: the alkene carbon farther from the alcohol forms the new C–C bond, placing a carbocation at the nearer alkene carbon. Trifluoroacetate captures that cation; the basic methanolic work-up cleaves the trifluoroacetate ester to reveal the ring hydroxyl group. This is a Prins cyclization.
Number the product ring from oxygen through the aldehyde-derived carbon as C2, then the alkene-derived methyl-bearing carbon C3, the hydroxyl-bearing carbon C4, the intervening methylene C5 and the original alcohol carbon C6. A chair keeping the bulky C2 side chain and C6 hydroxymethyl group equatorial predicts 2,6-cis substitution. Suprafacial alkene capture preserves the E-alkene-derived relationship, giving 2,3-trans between the C2 side chain and C3 methyl. Preferred equatorial trapping of the C4 cation gives 3,4-trans, hence the 2,4,6-cis arrangement of the substituent-bearing bonds in the favoured chair. In the printed flat-ring orientation, draw the C2 side chain, C4 hydroxyl and C6 hydroxymethyl behind the plane, and C3 methyl in front. Existing stereocentres in the aldehyde side chain and at C6 are retained; they anchor the absolute face of this assignment.

Articles by others on the same topic
There are currently no matching articles.