D is racemic trans-2-methylcyclohexan-1-ol. E is racemic 1-methyl-2-phenylcyclohexan-1-ol with OH and Ph trans, and Me and Ph cis. Number the starting alkene carbon carrying methyl as and its partner as in the reaction explanation below; the product name of D renumbers the OH-bearing carbon as 1.
For D, hydroboration proceeds through a concerted four-centre transition state: the alkene pi pair donates to boron while B–H supplies the new C–H bond. Boron attaches mainly to less substituted and hydrogen to , on the same face. Under peroxide/base conditions, hydroperoxide attacks B; migration of the C–B bond to the adjacent oxygen, with O–O cleavage, replaces C–B by C–O with retention at carbon. Hydrolysis/proton transfer releases the alcohol. Thus the newly added H and OH are syn, making the pre-existing methyl and OH trans. Attack on either alkene face gives the enantiomeric pair.
For E, the peracid transfers its terminal oxygen across the alkene in concerted epoxidation, through cyclic electron flow that also breaks O–O and transfers the acidic proton to the peracid carbonyl oxygen. The resulting epoxide retains the relative configuration of the alkene substituents. The phenyl equivalent from phenylmagnesium bromide then attacks less substituted from the face opposite the epoxide oxygen. Its C–O bond breaks onto oxygen, an SN2-like ring opening; the oxygen stays attached to . Water workup protonates that alkoxide. Consequently OH at and Ph at are anti, while the methyl at lies on the same relative face as Ph.
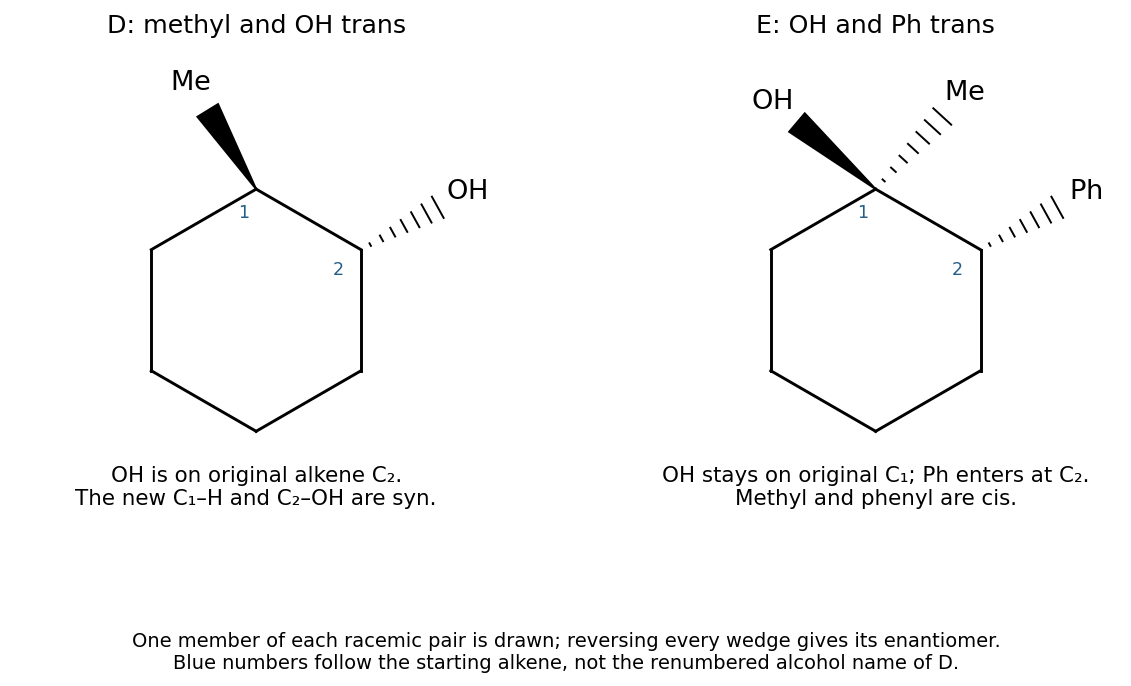
Original product structures showing the relative stereochemistry of hydroboration–oxidation and epoxide opening
. Details. 
Articles by others on the same topic
There are currently no matching articles.