Student suggested answers for the past ten years are available on Chemistry Moodle. Official course guide: past papers and suggested answers New solutions for years covered by that archive are skipped; existing wiki drafts are retained.
The held 2015 IA paper is also skipped: the 2023/24 IA handbook advertises suggested answers for papers more than five years old.
The diagnostic infrared spectroscopy absorptions are the conjugated ester carbonyl near and the alkene stretch near . Reasonable approximate ranges are – and –, respectively. Conjugated system lowers the carbonyl stretching spectroscopic wavenumber relative to an unconjugated ester.
The ester C–O stretches and strong C–F stretches lie mainly in the – region. Vinylic C–H stretching is just above , and saturated C–H stretching is about –. There is no hydroxy group to give a broad O–H band.
The assignments are : ppm, H, doublet of quartets; : ppm, H, doublet of quartets; : ppm, H, quartet; : ppm, H, triplet. The chemical shift of a multiplet is its centre, rather than its most downfield line.
Each vinylic proton has scalar spin-spin coupling to the other vinylic proton and to three equivalent spin- fluorine nuclei. The first interaction produces a doublet, and the second a quartet: eight lines overall, partly interleaved for . The two chemical shifts differ because the vinylic sites have different substituents.
The ethoxy protons are deshielded by oxygen and split into a quartet by the three methyl protons. The methyl protons are split into a triplet by the two equivalent methylene protons. The relative NMR signal integrals are .
Use the NMR coupling constant conversion . At , separating the overlapping quartets givesThe doublet separation independently gives . For , the two quartets are and ppm; simply taking adjacent lines in their combined ordering would mix different scalar spin-spin couplings.
The methylene quartet gives an average adjacent spacing ppm, or . Thus its printed rounded positions agree with the methyl-derived value to about ; quote the ethyl NMR coupling constant as approximately –. The large vinylic value supports the trans arrangement. The arrows below join the interacting nuclei; each arrow to represents the same interaction with each equivalent fluorine nucleus.
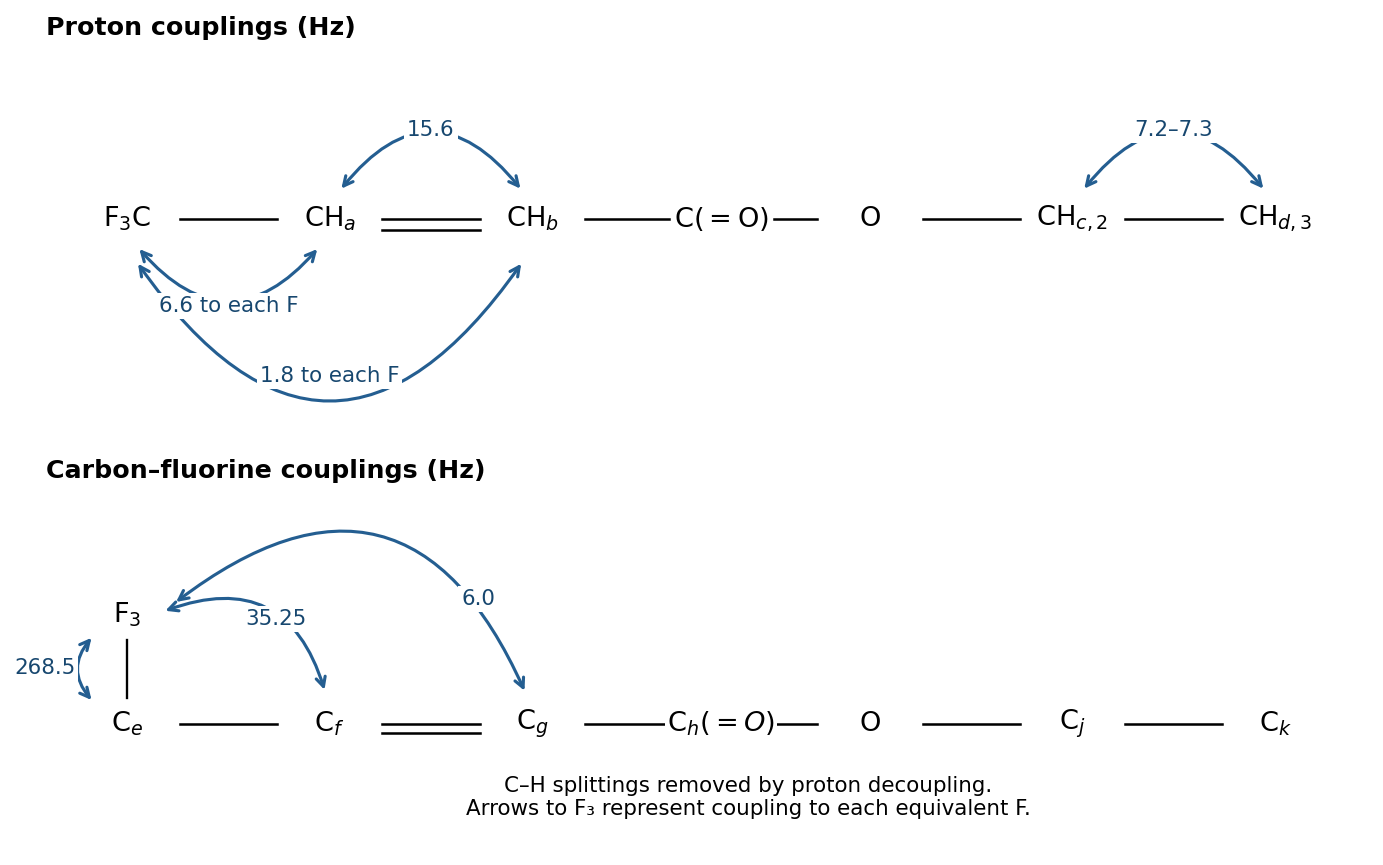
Calculated scalar spin-spin couplings for the vinylic, ethyl and trifluoromethyl sites
. Details. The six carbon environments are assigned as follows, with chemical shifts at the centres of the fluorine-split multiplets:The supplied carbon-13 spectrum is interpreted as proton-decoupled NMR, as indicated by its unsplit ethoxy signals. Proton decoupling removes C–H splitting but leaves C–F scalar spin-spin coupling. Three equivalent spin- fluorine nuclei give each resolved quartet its intensity pattern. The carbonyl's much smaller four-bond C–F interaction is unresolved in the plotted spectrum; “singlet” describes what is observed, rather than proving an exactly zero interaction.
The wide quartet is centred at ppm. The other two centres are and ppm. The signal near ppm is the deuterated chloroform solvent, not another carbon of the analyte. The structure labels skip : the ethoxy methylene is .
At a carbon-13 resonance frequency of , the three resolved NMR coupling constants areEach is the spacing of adjacent lines within its own quartet. The lower panel of the original coupling diagram in 1(b)(ii) marks the corresponding carbon–fluorine interactions. The bond superscripts count bonds along the connecting path; they are not additional numerical factors multiplying .
One fluorine-19 NMR resonance, integrating to three equivalent fluorine nuclei, appears as a doublet of doublets with and . This predicts four principal lines of equal intensity, from coupling to and , respectively. Relative to the unknown resonance centre, their offsets are and .
Equivalent fluorine nuclei do not split one another. This prediction assumes ordinary proton-coupled acquisition; proton decoupling would collapse these four principal lines to a singlet. The proton and carbon spectra do not determine the numerical fluorine chemical shift. Weak natural-abundance carbon-13 satellites could accompany the principal pattern.
For a hydrogen-like ion, the Coulomb potential is stronger by the nuclear atomic number , and the energy levels scale as :Here is the number of protons in the atomic nucleus; the ion has exactly one electron, so its net electric charge is . The zero of energy is a separated stationary electron and atomic nucleus at infinite distance. This is the nonrelativistic hydrogenic approximation using the supplied ; a more precise isotope-specific result also corrects the reduced mass.
The lowest two energy levels have and . For carbon, , even though the ion's net charge is :Increasing raises the energy toward zero; the most negative value is the ground-state energy.
An emitted photon has positive energyFor spontaneous emission into , the conservation of energy equation isConsequentlyThe principal quantum number must be an integer, so the transition is . Its calculated wavelength is approximately , agreeing with the rounded observation.
End-on overlap produces and molecular orbitals; side-on overlap produces and molecular orbitals. Matching signs of the facing atomic orbital lobes give constructive interference and a bonding molecular orbital. Opposite signs give destructive interference and an additional nodal plane between the atomic nuclei in the antibonding molecular orbital.
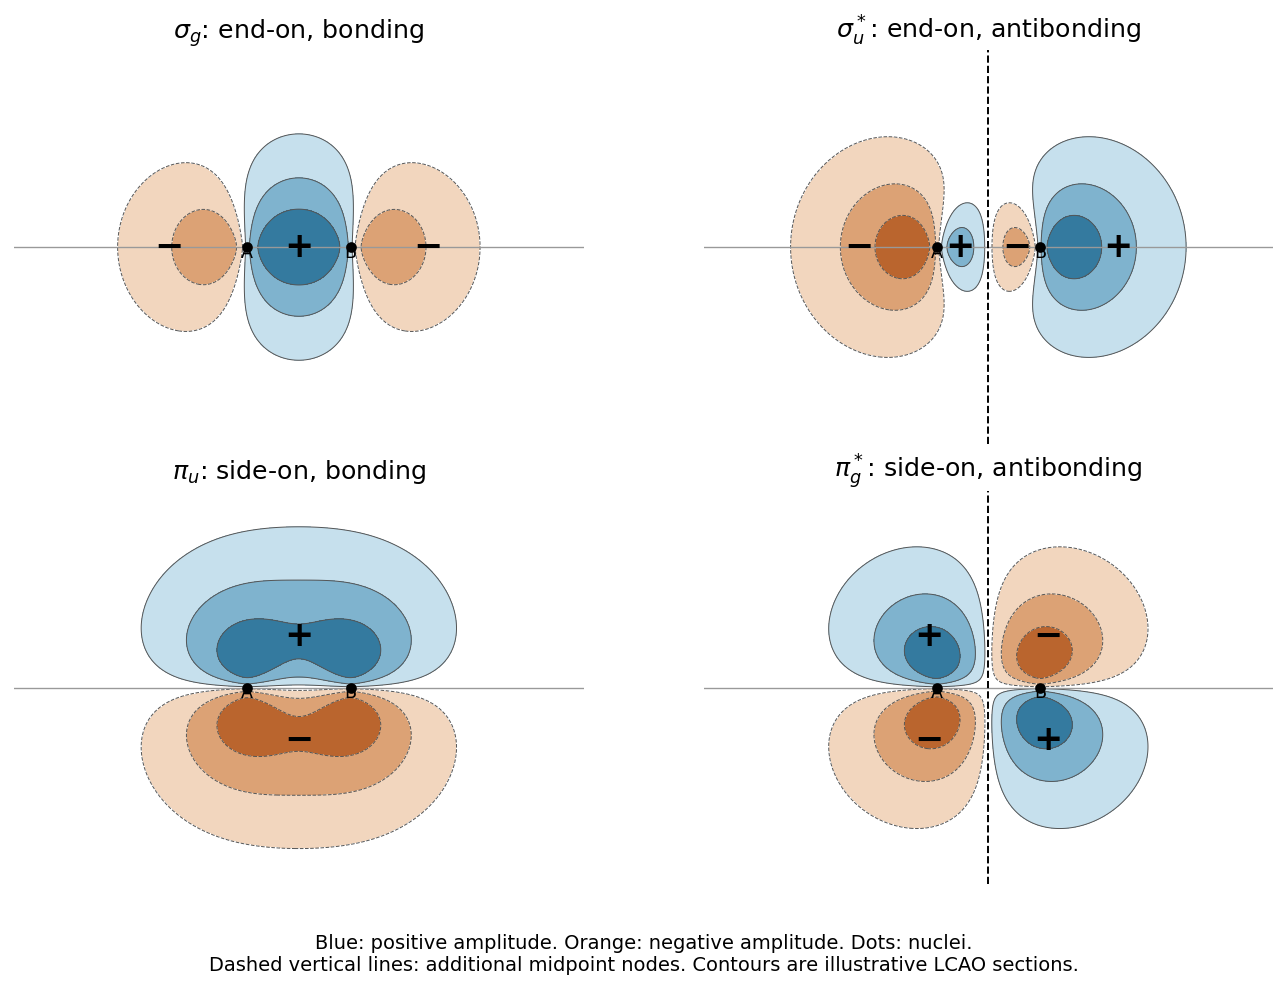
Take the internuclear direction as , and define both atomic functions as positive on their own right-hand side. Up to wavefunction normalization, the combinations areThe minus sign in the bonding combination follows from this common-axis quantum phase convention: the facing lobes of the two unmodified functions have opposite signs. It does not make that combination antibonding. Every molecular orbital also has a nodal plane containing the internuclear axis; the additional midpoint node distinguishes from .
The labels apply when the molecule has an inversion centre, as a homonuclear diatomic molecule does. The label means , and means , with the origin at that centre. In a heteronuclear diatomic molecule, inversion exchanges different atomic nuclei and these are generally not valid symmetry labels.
For the p-derived combinations above,Inversion both exchanges the two centres and changes the sign of each individual p atomic orbital. Applying both operations to each sum or difference gives these labels. The star denotes antibonding molecular orbitals, independently of parity: a bonding molecular orbital need not always be .
One consistent localized orbital hybridization description uses the Lewis structuresFor methyldiazonium, take C as and both N atoms as . For diazomethane, the double-bond resonance contributor suggests C as , central N as , and terminal N as . The central N uses two opposite hybrids for the linear C–N–N sigma bond framework. In the localized double-bond picture, terminal N uses one hybrid for its sigma bond, two for lone pairs, and its remaining p atomic orbital for a pi bond.
These labels specify an approximate bonding construction, not uniquely measurable atomic properties. An alternative representation takes terminal N as , with one lone pair and one p lone pair; the latter can participate in the delocalized C–N–N system. It describes the same resonance more explicitly. Neither construction requires an unpaired electron or a bent central N.
There are valence electrons, all paired, so diazomethane exhibits diamagnetism. A localized orbital occupancy consistent with the double-bond construction in 2(c)(i) isHere mean central and terminal. Each listed occupied spatial orbital contains two electrons of opposite spin. The central N's two p atomic orbitals are perpendicular, so they can support the two perpendicular pi bonds. No central-N lone pair is left over: its positive formal charge accompanies four bonding pairs in this Lewis structure.
Allowing resonance mixes a terminal-N lone pair into the C–N–N orbitals. In the alternative terminal- basis, the same electron count is four occupied orbitals ( electrons), one terminal- lone pair (), one perpendicular N–N orbital (), and two occupied orbitals of a three-centre C–N–N system (). The -electron count and the paired ground state survive this change of description; the localized picture does not assert two perfectly isolated double bonds.
On carbon protonation, the C–N bond length increases and the N–N bond length decreases. Forming the fourth C sigma bond changes the carbon from approximately planar to tetrahedral , removing its effective C–N pi bonding. The product is described by : C–N becomes predominantly single and N–N predominantly triple. The resonance of the unprotonated species makes its actual bond orders intermediate rather than exact integers.
The products are hexanenitrile, , and ethyl triphenylmethyl ether, . The nitrile carbon comes from cyanide, so replacing chloride in the five-carbon chain gives a six-carbon product.
For the primary chloride, the carbon-end lone pair of attacks the carbon carrying Cl from the back side while the C–Cl bond pair moves onto Cl. The single concerted nucleophilic substitution step forms the C–C bond and releases . Attack through the carbon end gives the nitrile, not an isocyanide.
For the triphenylmethyl chloride, the C–Cl bond first undergoes heterolysis to and . An ethanol oxygen lone pair then attacks the carbocation, giving . Finally pyridine accepts that O–H proton, with the O–H bond pair returning to oxygen, giving the neutral ether and pyridinium chloride.
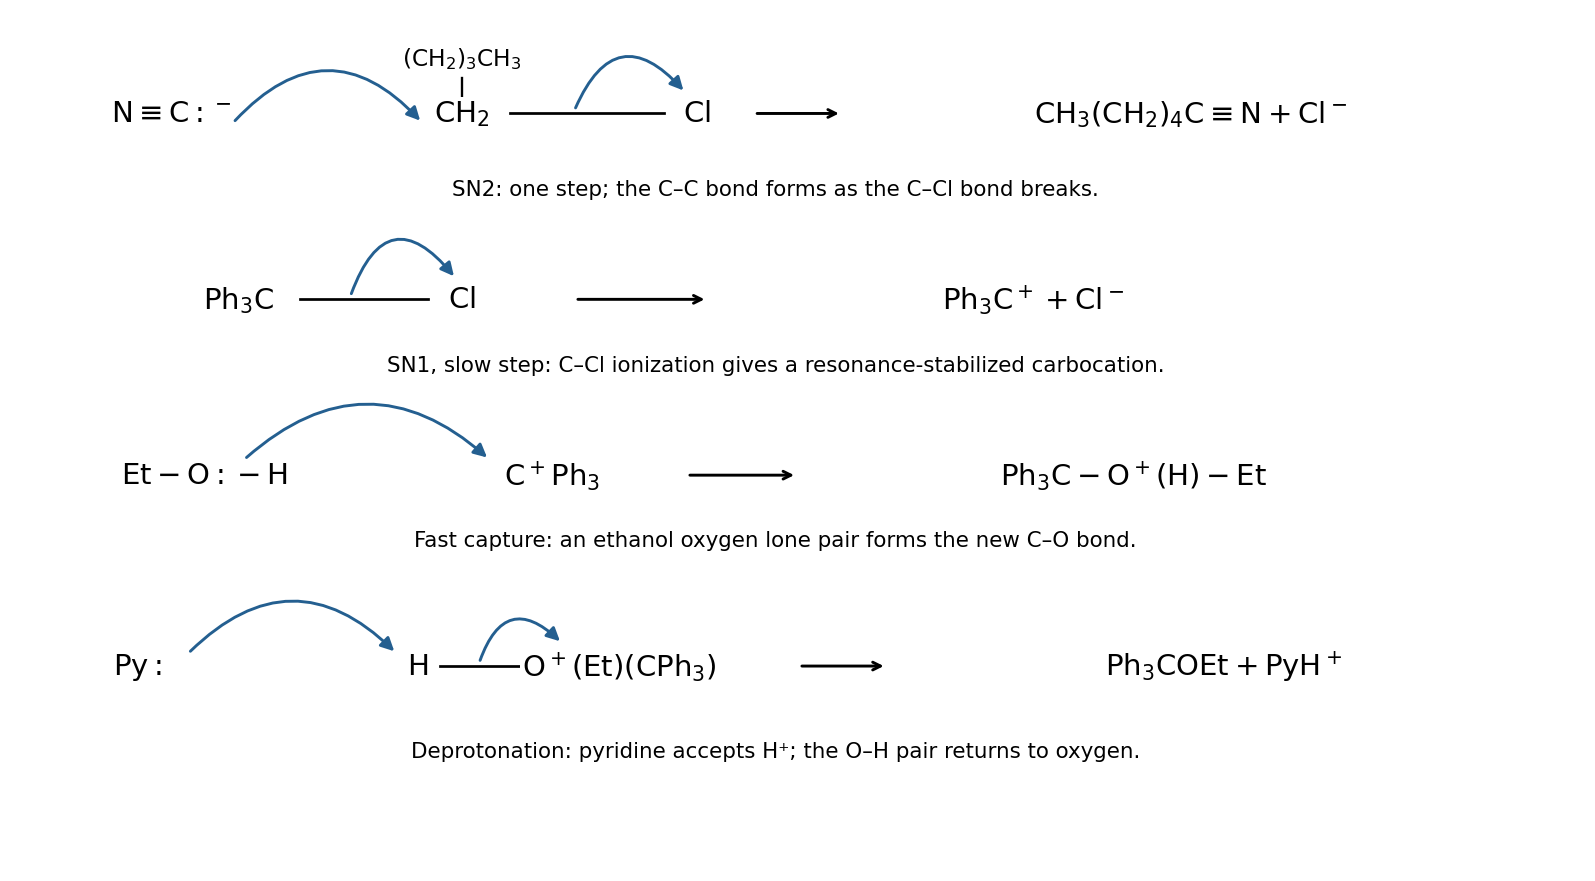
The primary chloride undergoes an SN2 reaction; the triphenylmethyl chloride undergoes an SN1 reaction. The primary site is accessible to back-side attack, whereas three phenyl groups make back-side attack at the triphenylmethyl site extremely hindered and stabilize its carbocation through resonance.
For the SN2 reaction, a suitable transition state isThe entering C and leaving Cl lie on opposite sides of the reacting carbon, with partial C–C and C–Cl bonds. The total charge is . This transition state is not a discrete reaction intermediate.
For the SN1 reaction, the approximately planar carbocation is a genuine reaction intermediate. Assuming its formation by C–Cl heterolysis is rate determining and trapping is rapid,Thus the limiting rate law is independent of ethanol concentration. The successive attack and proton transfer occur after the slow ionization; a cation-stabilizing environment and efficient trapping underpin this idealized SN1 description.
B is 4-hydroxy-1-methylpyrrolidin-2-one; C is 1-methylpyrrolidin-3-ol. Their ring connectivities are shown below. Each ring remains five membered; the attached N-methyl group is retained.
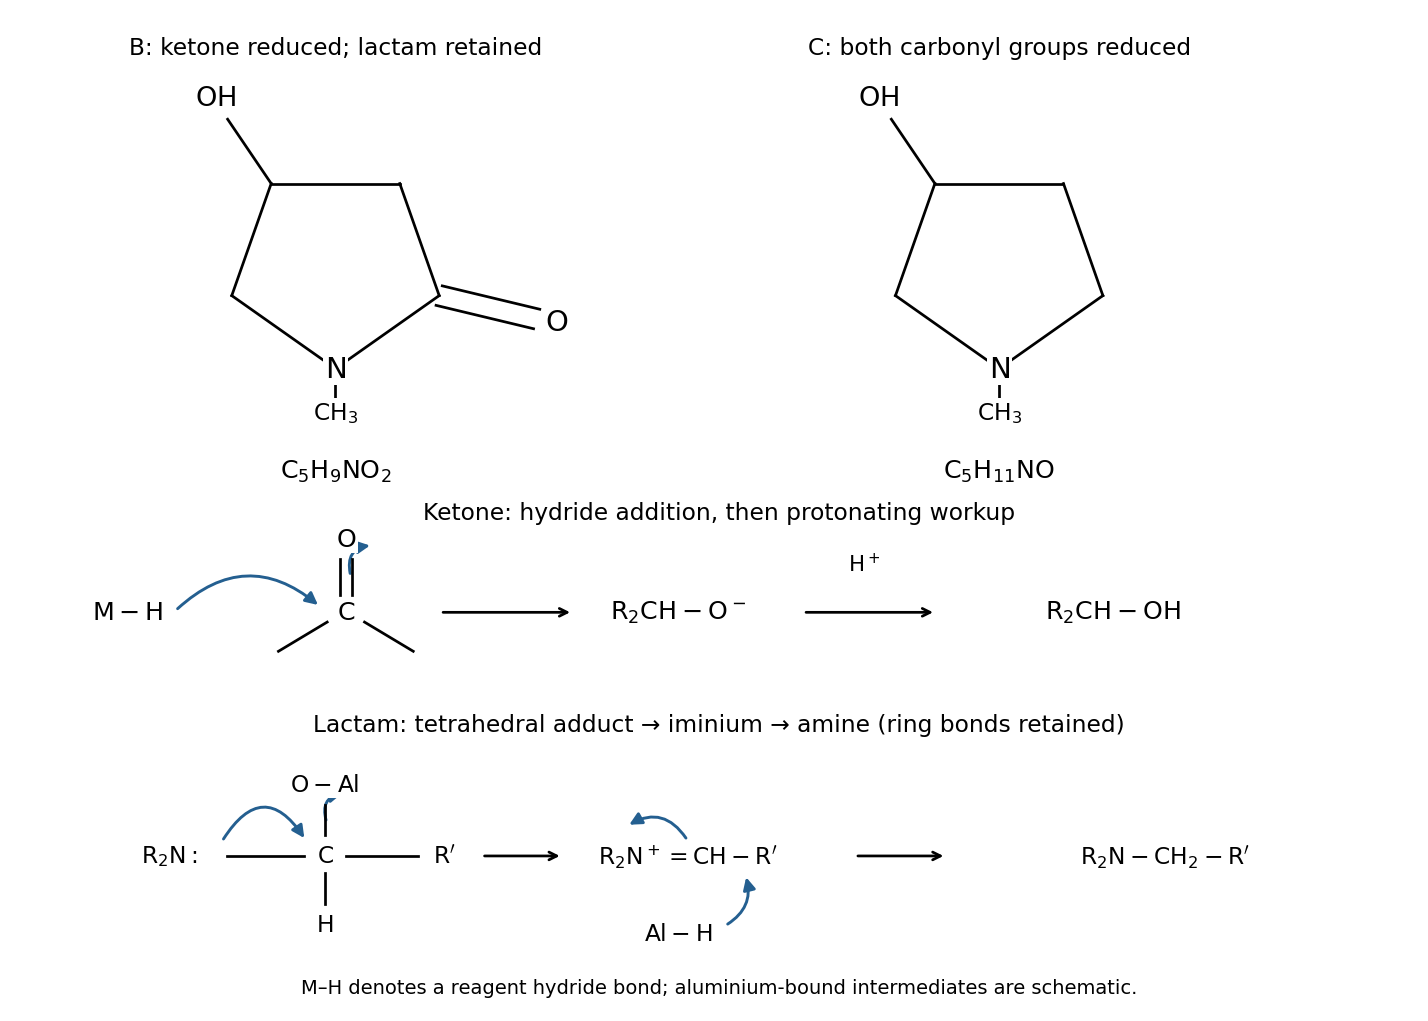
For B, a B–H bond of sodium borohydride supplies a hydride equivalent to the ketone carbonyl carbon, and the carbonyl pair moves to oxygen. This gives an alkoxide, whose protonation during the reaction or workup gives the secondary alcohol. The lactam carbonyl remains. Converting one to adds two H atoms and gives .
For C, lithium aluminium hydride reduces that ketone in the same net way and also reduces the lactam. Coordination of the amide oxygen to aluminium assists hydride attack at its carbonyl carbon, giving a tetrahedral intermediate. The N lone pair then forms a C=N bond while the C–O bond breaks to an aluminium-bound oxygen group. A resulting iminium ion accepts a second hydride at carbon, yielding the cyclic tertiary amine. Aqueous workup protonates the ketone-derived alkoxide.
The lactam conversion replaces by ; its oxygen is removed rather than retained as another alcohol. Together with ketone reduction, the net change is , giving . No stereochemistry is specified: the new alcohol stereocentre is formed as a racemic mixture in an achiral environment.
Sodium borohydride is chemoselective for the ketone; lithium aluminium hydride reduces both the ketone and the lactam. In an amide, donation of the N lone pair into the carbonyl system reduces the carbonyl carbon's electrophilicity. It is therefore substantially less susceptible to the milder hydride donor.
Lithium aluminium hydride supplies more strongly reducing hydride equivalents, and the strong affinity of aluminium for oxygen assists removal of the amide oxygen through the iminium pathway. This difference explains both the unchanged lactam carbonyl in B and the absence of that oxygen in C. Both reactions are interpreted after the usual protonating workup.
D is , and E is . D results from transesterification; E results from acid-catalysed acetal hydrolysis.
For D, methoxide attacks the ester carbonyl carbon and moves the carbonyl pair to oxygen. Collapse of the tetrahedral reaction intermediate reforms C=O and expels ethoxide. Proton transfer with the methanol solvent regenerates methoxide. Excess methanol favours replacement of the ethyl ester by a methyl ester; the dimethyl acetal survives these basic conditions.
For E, protonate one acetal oxygen, then break its C–O bond with the pair going to oxygen, releasing methanol and forming an oxocarbenium ion. Water attacks that carbon. Deprotonation gives a hemiacetal; protonation of its remaining methoxy oxygen permits loss of a second methanol. The hydroxyl oxygen's lone pair forms C=O during this loss, and deprotonation gives the aldehyde, regenerating the acid catalyst.
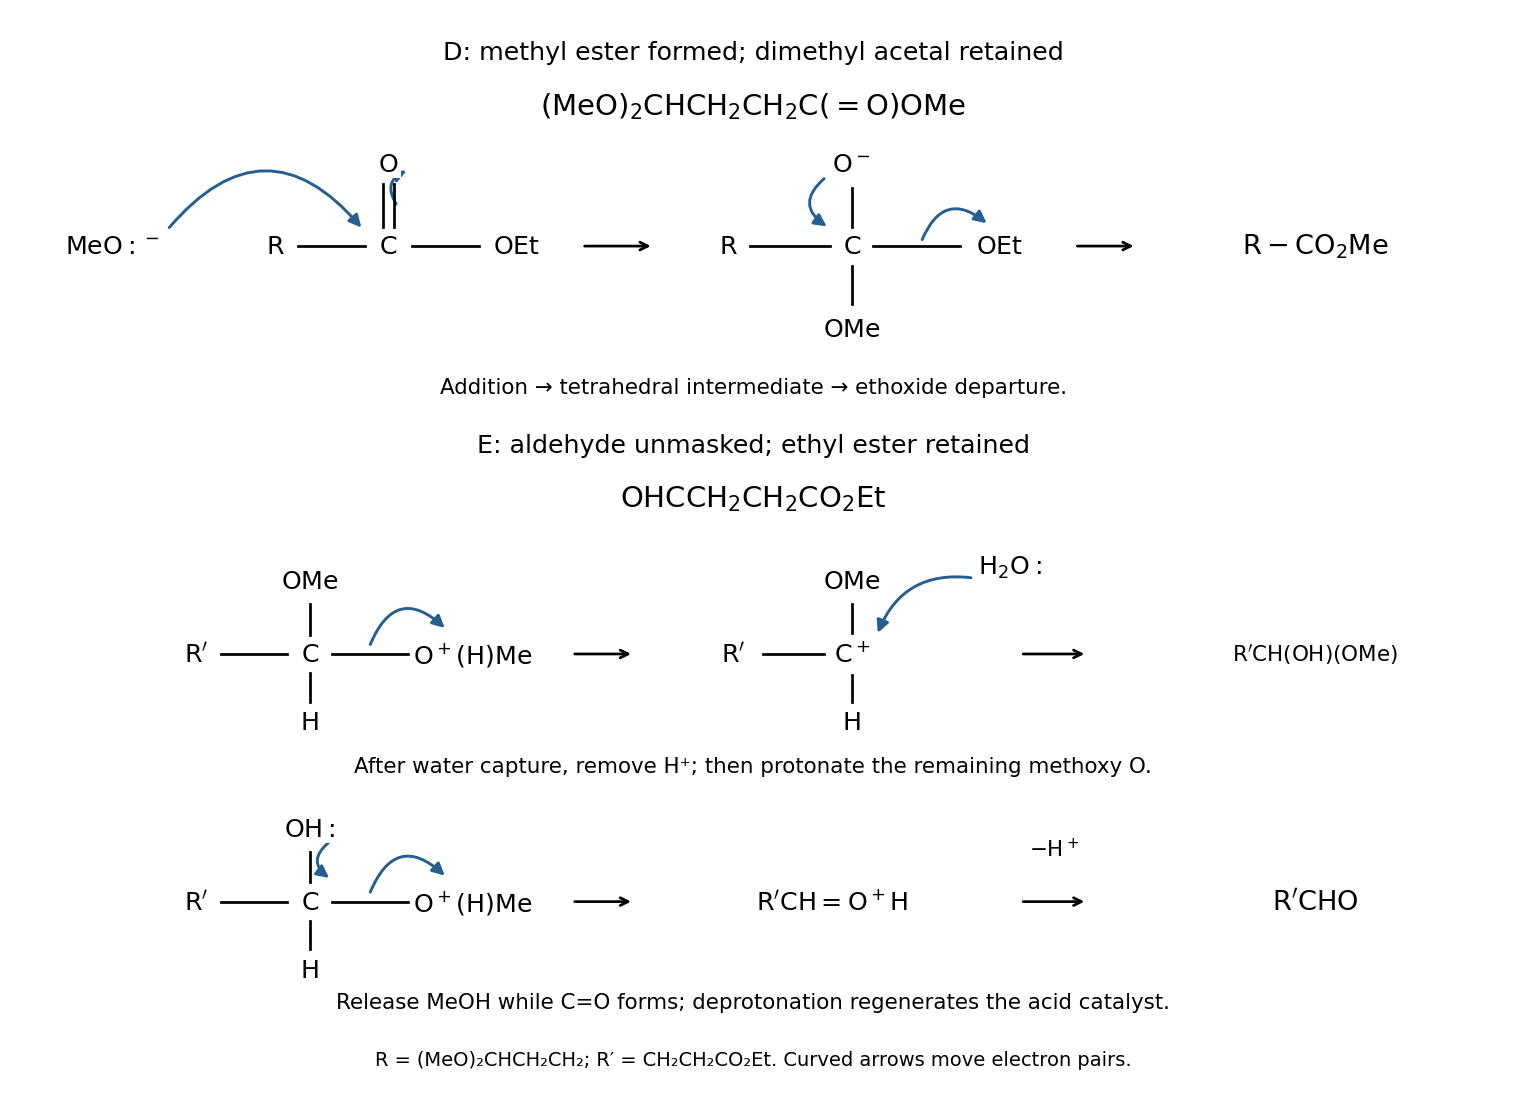
Original products and electron-flow steps for transesterification and acid-catalysed acetal hydrolysis
. Details. The spectroscopy is consistent with these products. D has one ester carbonyl near and six carbon-13 NMR environments: acetal CH, two distinct chain sites, carbonyl, ester , and the two equivalent acetal groups. Thus seven carbon atoms give six resonances. E has separate ester and aldehyde carbonyl bands, approximately and , and six distinct carbon atoms: two carbonyls, two chain methylenes and the two ethyl carbons.
Basic methanol exchanges the ester alkoxy group while retaining the acetal; dilute aqueous acid removes the acetal while retaining the ethyl ester on the stated reaction timescale. An acetal lacks an electrophilic carbonyl and is stable to ordinary base: expelling methoxide directly would require a poor leaving group. The ester can instead undergo nucleophilic acyl substitution through a carbonyl-derived tetrahedral intermediate.
Under acid, protonating an acetal oxygen converts an alkoxy substituent into departing methanol, and the resulting oxocarbenium is stabilized by its remaining oxygen. Water then drives acetal hydrolysis toward the aldehyde. Ester hydrolysis is also possible under acid, but is slower under the conditions implied by the product data. The retained ethyl group and six distinct carbon-13 NMR environments support E rather than the carboxylic acid that prolonged hydrolysis could produce.
With ideal thermodynamic activities , the dimensionless acid dissociation constant isThe factor makes the equilibrium constant compatible with . If the concentration quotient without is used instead, it has concentration units and its numerical value is identical only when the standard concentration is expressed as .
The standard Gibbs energy of reaction givesThe positive standard Gibbs energy of reaction means that dissociation is unfavourable when every thermodynamic activity equals one; it does not forbid partial dissociation. The corresponding is about .
The equilibrium molecular-acid concentration is . From the supplied pH convention,With only these monovalent ions appreciably present, electroneutrality requires . Substitution in the acid dissociation constant givesThis calculation assumes ideal thermodynamic activities for the acid and its conjugate base as well as for , negligible water autoionization, negligible second dissociation, and no additional electrolyte. The acid's analytical concentration would include its dissociated fraction and be approximately , rather than exactly the molecular-acid concentration requested.
Oxidize zinc at the anode and reduce nitrate to nitric oxide at the inert platinum cathode. Multiplying the oxidation half-reaction by three and the reduction half-reaction by two cancels six electrons:The standard cell potential is the cathode reduction potential minus the anode reduction potential:The written direction is spontaneous under standard conditions, since . Multiplying a half-reaction changes its extensive Gibbs energy, but does not multiply its intensive electrode potential.
The Nernst equation for the six-electron cell reaction isHere pure solid zinc and liquid water have unit thermodynamic activity. For ideal nitric oxide gas, with ; for ideal dissolved species, . The reaction quotient is dimensionless. The platinum supplies an electronically conducting surface and is absent from the net chemical equation.
Bulk electroneutrality gives . Each dissolved zinc ion contributes two positive elementary charges, whereas each nitrate contributes one negative charge.
This assumes complete nitric acid dissociation, only and as significant cations, and only nitrate as a significant anion. It neglects at acidic pH, other dissolved salts, zinc complexes and additional ionic nitrogen products. It is a bulk charge balance; a thin charged electrode interface does not invalidate bulk electroneutrality.
At pH , , so the nitrate concentration is . With ideal thermodynamic activities,HenceThe activity of nitric oxide is because the supplied pressure is and the gas standard is . This numerical result additionally approximates zinc and nitrate thermodynamic activities by their standard-concentration ratios; without their activity coefficients, a more precise nonideal value is not specified.
For the cell reaction normalized as in 4(b)(i), six moles of electrons are transferred per mole of reaction. ThereforeThis is the Gibbs energy of reaction at the stated composition and gas pressure, not the standard Gibbs energy of reaction. Its negative sign confirms thermodynamic favourability in the written direction under those conditions.
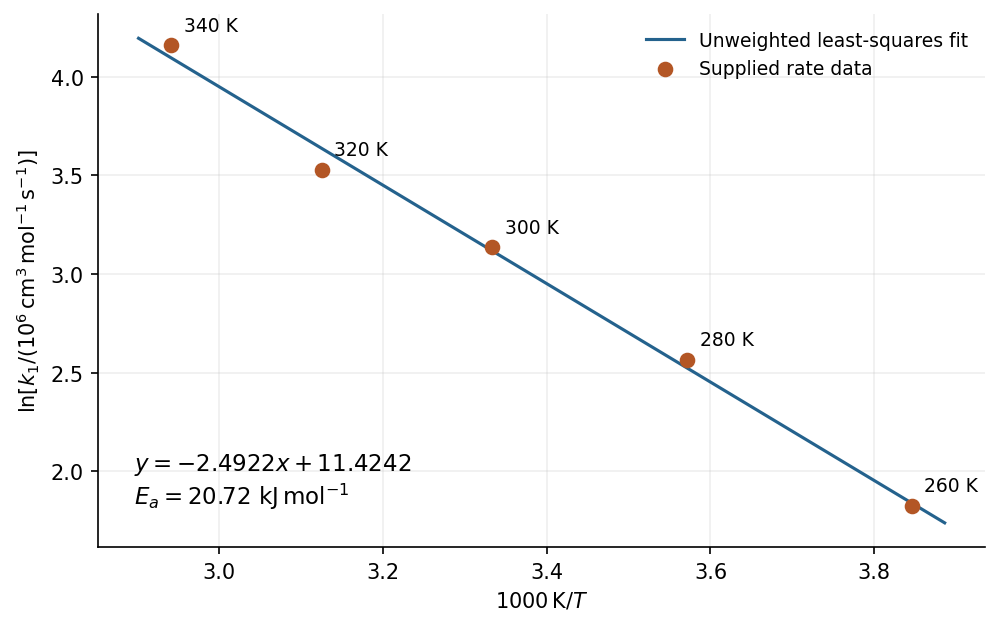
The Arrhenius equation isHere is the molar activation energy, the molar gas constant, the absolute temperature, and the pre-exponential factor, taken approximately constant over the measured range. For this bimolecular elementary reaction, and both have units .
An unweighted least-squares Arrhenius plot using all five data points gives . Define the dimensionless ordinate with , and set . The model isFor increasing temperatures, the five ordinates are , , , , and . The linear regression slope and intercept areConsequently . The corresponding fitted . A two-point calculation using just the endpoints gives approximately ; the difference reflects the rounded, imperfectly collinear data rather than a unit conversion.


This labelled part introduces the reversible reaction pair without a separate calculation to perform. For the following derivations, define its forward and reverse elementary reaction rates asand the initiating rate as . The law of mass action gives bimolecular units and first-order units. Reversibility does not require while the complete network is reacting: that equality would impose chemical equilibrium on the pair, an extra assumption not needed here.
The quasi-steady-state approximation for the nitrate radical balances its formation against its removal:Equivalently,The net dinitrogen pentoxide production is . Substituting the steady-state balance givesThe reverse rate has been retained throughout and cancels algebraically. The approximation requires sufficiently rapid radical adjustment and fails if, for example, nitrogen dioxide becomes too scarce to sustain the presumed trapping rate. It does not assume that the radical concentration is zero.
Nitrogen dioxide is consumed in reactions 1 and 2 and regenerated in reaction , soUsing from the nitrate-radical quasi-steady-state approximation givesThe factor of two is essential: an initiating event uses one nitrogen dioxide molecule, and the ensuing net radical-trapping event uses a second one. The exact nitrogen balance before making this approximation is .
Adding the initiation step to the net forward trapping step cancels the nitrate-radical reaction intermediate:The net stoichiometry agrees with under the quasi-steady-state approximation.
The remaining fraction is . Using the tabulated value rather than replacing it by the fitted value, the constant ozone concentration makes nitrogen dioxide loss a pseudo-first-order reaction:Integrating with initial concentration and givesThis uses the nitrate-radical quasi-steady-state approximation over the interval, constant temperature and ozone concentration, and no extra nitrogen dioxide sources or sinks. The stated and units cancel directly; no factor of is needed.
For the outer valence electrons, Slater's rules predict a modest initial rise down group 15, followed by approximately constant values. Write the effective nuclear charge as , where is the shielding constant. Each atom has , so the other four electrons in the outer group contribute . Electrons in shell contribute each, and those deeper contribute each.[ref]
Atomic Shielding Constants by John C. Slater. 1930, shielding rules on printed pp. 58–59
Applying those weights gives the following original calculations:The increasing nuclear electric charge is largely offset by additional inner-electron shielding; once the preceding shell contains electrons, these simple weights give a plateau. The shell includes its d electrons. This trend concerns outer-shell values, not every orbital: a electron is shielded only by its partner, so rises strongly down the group. The larger principal shell also means nearly constant outer does not imply nearly constant atomic size or binding energy.
Photoelectron spectroscopy measures orbital-dependent binding energies from which an effective charge can be inferred using a stated model. Ultraviolet photoelectron spectroscopy accesses outer-shell ionizations; X-ray photoelectron spectroscopy accesses deeper core states. For an isolated gas-phase atom and a calibrated threshold, measure emitted-electron kinetic energy and use , with the appropriate energy reference. Optical atomic term spectra and series limits also constrain valence ionization energies.
A hydrogenic inference would set , hence , with here an energy of approximately and a chosen effective shell parameter. The measured binding energies are experimental; the extracted values remain model dependent. They need not equal Slater screening parameters fitted to radial functions.
Unlike the equal values in Slater's rules, spectroscopic values distinguish s from p because their penetration and shielding differ; the more tightly bound s states usually imply larger binding-energy-derived effective charges at a common shell parameter. Subshell, atomic-term and, especially for heavier atoms, spin-orbit coupling structure give further distinctions. Electron relaxation following ionization and electron correlation also matter. Thus real binding energies vary across the heavier group-15 atoms even where the simple Slater outer-shell estimates plateau; no universal constant correction to the Slater values follows.[ref]
Electron Spectroscopy for Atoms, Molecules and Condensed Matter by Kai Manne Börje Siegbahn. 1981, binding-energy measurements and resolved atomic and chemical environments
Phosphorus has electron configuration , with . For a electron, the only shielding contribution in Slater's rules is from the other electron:For a electron, the other four electrons in the group contribute , the eight electrons contribute , and the two electrons contribute :The compact core electron experiences nearly the full nuclear charge, while the valence electron is strongly shielded by the core. The two numbers describe different orbitals of the same atom. Outer electrons do not shield the core in this Slater approximation, and the selected electron never counts toward its own shielding.
Allen electronegativity is proportional to the average binding energy of the free ground-state atom's valence electrons. For an s,p main-group atom, use positive, spectroscopically inferred one-electron binding energies and their occupancies :Here fixes the conventional scale; the one-electron energies are obtained from appropriate atomic ionization/term energies, not merely from the first ionization energy. For group 15 the weighted average is .[ref]
Electronegativity is the average one-electron energy of the valence-shell electrons in ground-state free atoms by Leland C. Allen. 1989, definition and spectroscopic construction
The approximate connection to effective nuclear charge is . At a fixed shell, stronger effective nuclear attraction generally raises the valence binding energies and therefore Allen electronegativity. Down a group, increasing shell size and changes in penetration offset that attraction. There is no universal relation across different shells, nor can one substitute a core-electron for the valence average.
Gallium phosphide forms a tetrahedral network of polar covalent bonds, with electron density displaced toward phosphorus. The alternating diamond-related structure is the zinc blende structure: each Ga has four P neighbours and each P has four Ga neighbours. A local orbital hybridization picture combines outward-pointing s,p-derived orbitals into four approximately bond directions; in the crystal these interactions form extended occupied bonding bands.
Since , P attracts shared electron density more strongly. Write to indicate an ionic contribution superposed on covalent bonds. Formal oxidation states Ga() and P() are electron-assignment labels, not proof of completely transferred charges or a purely ionic crystal.
Neither atom requires hybrids for these four bonds. The Ga shell is filled and relatively core-like; the relevant Ga valence orbitals are . P uses principally ; its empty orbitals are too high in energy to be a necessary bonding set. Detailed band calculations may include d-state mixing or core–valence effects, but that does not turn the tetrahedral bonding into an expanded-valence d-hybrid construction.
Articles were limited to the first 100 out of 504 total.
Articles by others on the same topic
There are currently no matching articles.