The products areB retains the terminal alkyne and replaces the tertiary propargylic chlorine with ethoxy. D replaces the benzylic chlorine while retaining the aromatic chlorine; its meta arrangement follows from the drawn starting material, rather than from four aromatic protons alone.
| Product | Proton shift / ppm | Assignment |
|---|---|---|
| B | 1.21, 3H triplet | Ethoxy terminal methyl, coupled to the adjacent two protons. |
| B | 1.46, 6H singlet | Two equivalent methyl groups on the substituted carbon, with no neighboring proton to split them. |
| B | 2.66, 1H singlet | Terminal alkyne proton. |
| B | 3.59, 2H quartet | Ethoxy methylene next to oxygen, coupled to its methyl group. |
| D | 1.18, 3H triplet; 3.58, 2H quartet | The ethoxy group, with a common approximately 7 Hz coupling. |
| D | 4.80, 2H singlet | Benzylic methylene attached to oxygen, with no vicinal aliphatic proton. |
| D | Two aromatic multiplet regions, 4H total | The four remaining ring protons of the disubstituted aromatic ring. |
For B, the strong band is the terminal sp C–H stretch, the band is aliphatic C–H stretching, and the weak band is the alkyne stretch. The terminal proton and alkyne band together distinguish this from an alcohol interpretation of the high-frequency absorption.
For D, is aromatic C–H stretching, is aliphatic C–H stretching, and the and bands are aromatic skeletal stretches. The listed spectrum is a selected peak list, so omission of a C–O band is not evidence that the ether bond is absent.
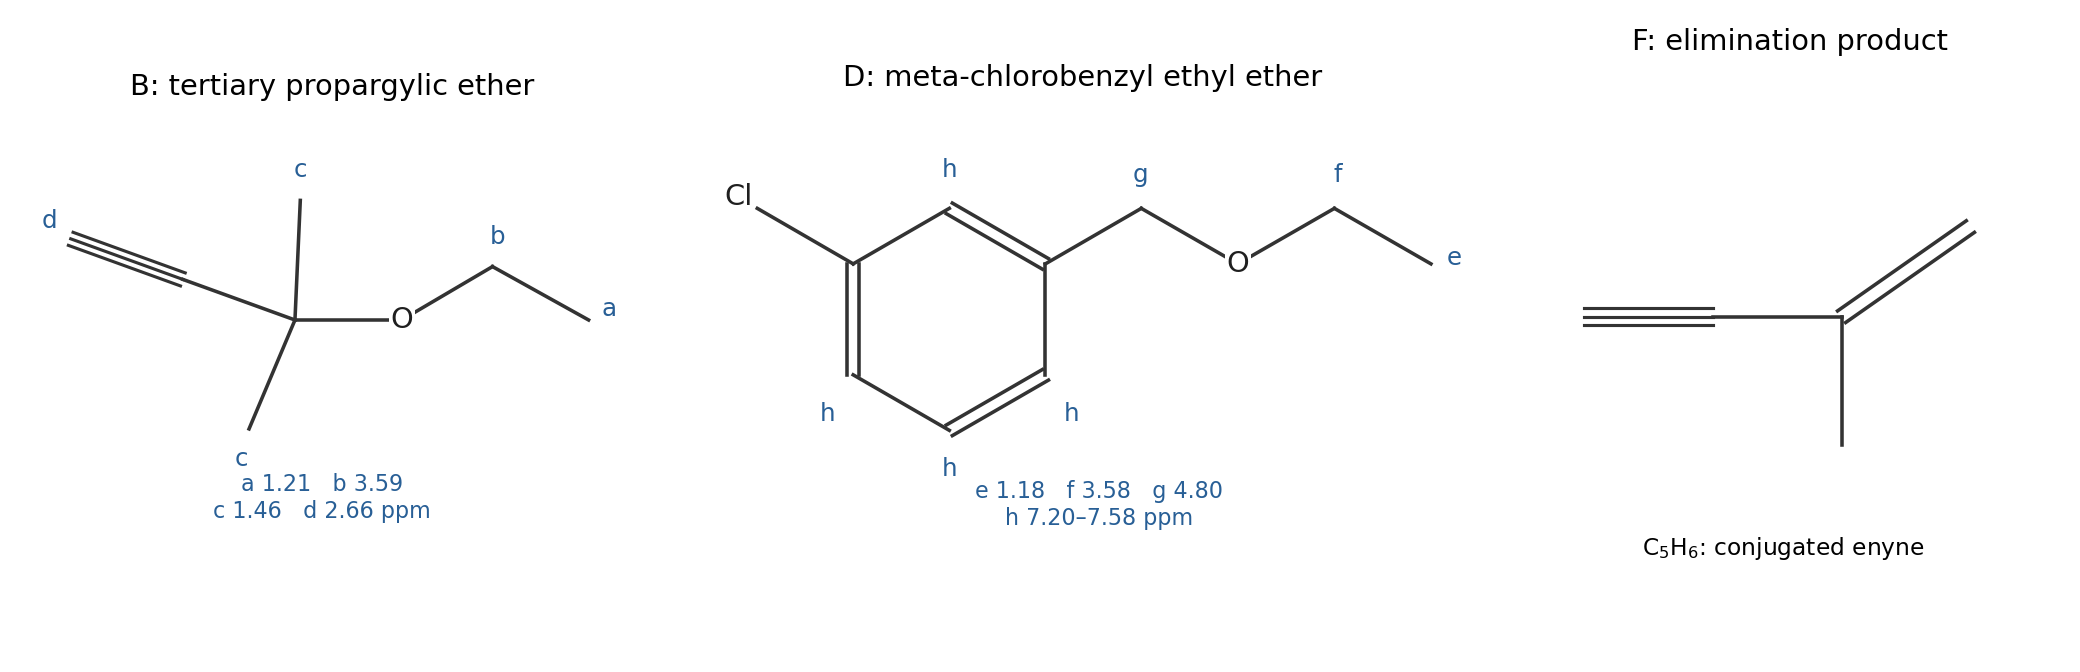
Derived ether products and the competing elimination product
. Original skeletal drawings of B, D and F. B and D retain the unsaturation indicated by the spectral data; F is the conjugated enyne formed by loss of hydrogen chloride from the tertiary propargylic substrate. Details.
For Reaction 1, doubling substrate concentration doubles the rate, while doubling ethanol concentration leaves it unchanged. ThusRate-limiting ionization of the tertiary propargylic chloride gives a carbocation stabilized by substitution and propargylic/allenylic resonance. The crowded reaction center disfavors backside attack, while neutral ethanol is a weaker nucleophile than ethoxide. Ethanol capture and deprotonation occur after the rate-determining step.
For Reaction 2, independently doubling either reactant doubles the rate, and doubling both gives a fourfold increase:The benzylic carbon is primary and accessible, and the strong nucleophile can displace chloride in a concerted step. The reacting bond is benzylic C–Cl, not the ring C–Cl bond. These kinetic orders support the proposed mechanisms in the stated model; the rate table alone would not identify every intermediate in a more complicated reaction network.
The drawn aryl dichloride is expected to undergo no analogous substitution under ordinary ethoxide conditions. Its chlorines are attached directly to sp2 aromatic carbons, rather than to the benzylic sp3 carbon of the previous substrate.
A normal SN2 reaction cannot follow the accessible backside geometry available at a primary benzylic carbon. A simple SN1 reaction would require a highly unfavorable aryl cation. Addition–elimination aromatic substitution would need suitable ring activation, such as a strong electron-withdrawing group in an appropriate position, which the drawing lacks. Forcing aryne chemistry cannot be inferred from ethoxide alone.
The conclusion addresses the actual E-plus-ethoxide scheme shown. The accompanying reference to replacing product B is inconsistent with the preceding substrate labels; interpreting it literally would not define the intended substitution comparison.
Elimination from A givesRemoving a beta hydrogen from either equivalent methyl group while losing chloride creates the same conjugated enyne. The strongly basic ethoxide, , promotes this E2 elimination from the tertiary substrate more strongly than neutral ethanol does. Ethanol instead favors the intended solvolytic capture, although some elimination can still compete.
The formula drawn and used in the rate table is ethoxide. The name “ethanoate” would mean acetate, , a different and resonance-stabilized anion; it should not be silently treated as the same base.
In Reaction 2 the primary benzylic carbon favors rapid substitution. The adjacent aromatic ipso carbon has no beta hydrogen for ordinary side-chain E2 elimination, so the analogous alkene-forming path is unavailable. This does not assert that all benzyl compounds are immune to every side reaction.
Articles by others on the same topic
There are currently no matching articles.