The ideal eclipsed ion has D4h molecular point group symmetry. Put the Mo–Mo axis along , with the metal atoms at , and orient the four chloride positions around each metal along .
There is a axis, perpendicular twofold axes, a horizontal plane through the midpoint, vertical mirror planes and an inversion centre at the bond midpoint. The eclipsed arrangement makes the two chloride squares coincide in projection along . These labels refer to the ideal molecular geometry, rather than small crystal-packing distortions.
Let and be corresponding real 4d orbitals on the upper and lower metals, with identical global-axis phases. Their symmetry-adapted linear combinations arewith the normalization shown in the negligible-overlap approximation; including overlap changes the denominator but not the symmetry. Inversion exchanges the centres and each local d orbital has even intrinsic parity, so the plus combinations are gerade and the minus combinations ungerade.The reversal of bonding phase for is important: the facing lobes have opposite signs when the two atomic orbitals share the same global phase convention, so their difference is bonding. Each irrep is two-dimensional. Thuswith total dimension ten. Rotating by interchanges the names of the two delta families; all labels here retain the chloride-aligned axes fixed above.
Ignoring ligand perturbations initially, end-on overlap gives the largest sigma bond splitting, side-on overlap gives the two pi bonds, and the two four-lobed overlaps give weaker delta bonds. A useful qualitative ordering isThe original diagram shows the ten orbitals, including both members of each level.
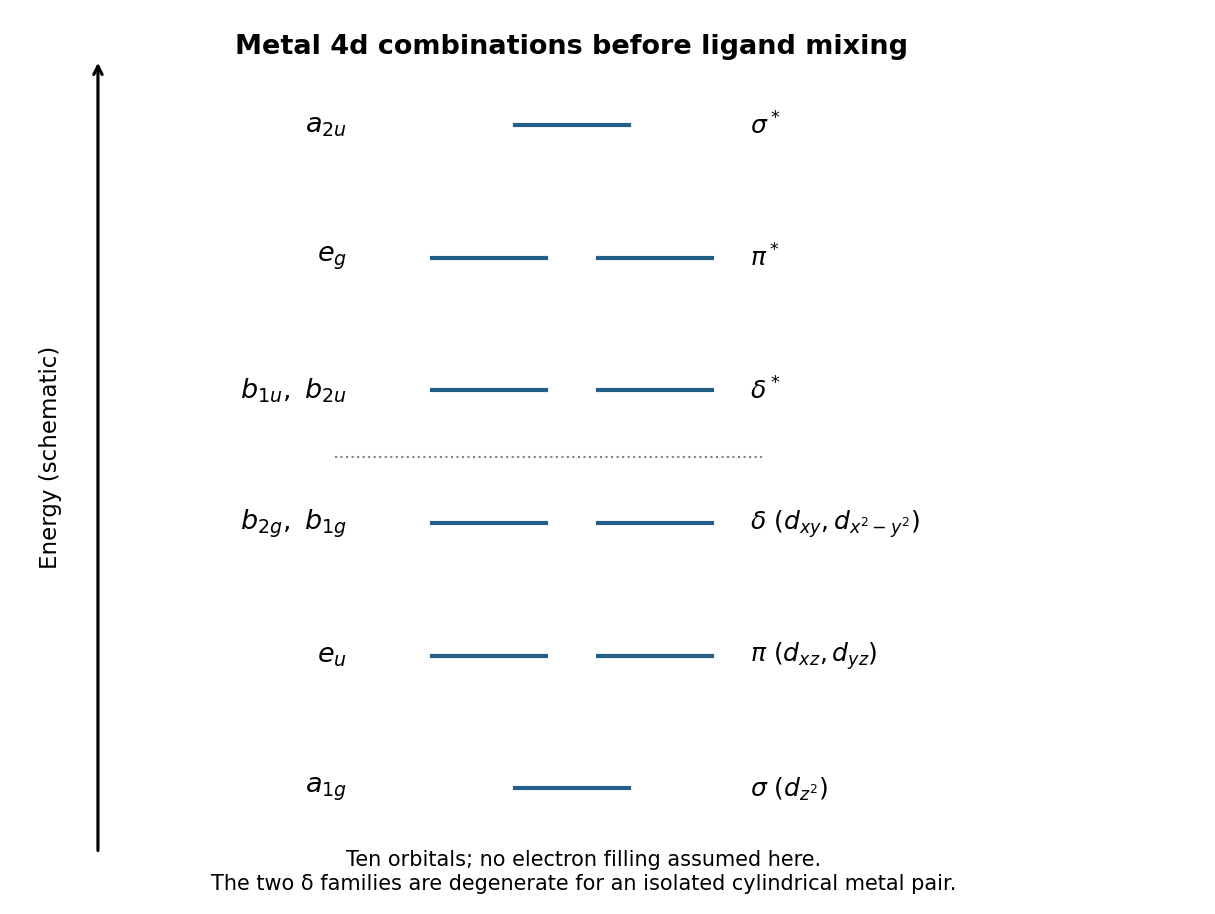
For the strictly isolated collinear metal pair, cylindrical symmetry makes and related by rotation, so the two delta bonding levels are degenerate, as are their antibonding partners. The displayed D4h molecular point group labels anticipate the ligand environment, which can split those different irreps. Symmetry supplies labels and degeneracies; the relative separation of unrelated levels is a qualitative overlap model, not a numerical prediction.
Eight chloride ions contribute charge . For an overall charge, the two metals together contribute , so each is formally Mo(II). Molybdenum is in group six, giving four d electrons per centre and eight metal-based electrons in total.
In the ligand-perturbed configuration developed below, they occupy , with no Mo–Mo antibonding electron. Therefore the formal bond order isThis is one sigma bond, two pi bonds and one delta bond. Before including ligands, the two ideal delta channels are degenerate and the allocation of their last two electrons is not determined by the bare energy sketch alone; the ligands remove that ambiguity and select the surviving low delta channel. The eight-electron bonding count still gives formal order four.
The occupied bonding groups are experimentally meaningful: molybdenum X-ray emission resolves three metal–metal bonding energy groups in this ion, alongside a deeper metal–chloride bonding feature.[ref]
Direct evidence for the quadruple metal-metal bond in the octachlorodimolybdenum(II) ion using X-ray emission spectroscopy by Derek E. Haycock and others. Primary X-ray emission report, abstract and pages 262–263
Choose the phase of each inward-pointing ligand sigma donor orbital positive toward its own Mo centre. The eight donors then transform by site permutation. In the usual D4h molecular point group class order their characters areOnly and a chloride-containing vertical mirror plane leave ligand functions individually fixed. For example fixes the four donors at and exchanges the other four; the four fixed functions retain their signs.
Molecular character reduction givesThe dimensions sum to . Equivalently, take each square's four combinations: the all-in-phase pattern, the alternating x-versus-y pattern, and a two-dimensional pair. Combine corresponding patterns between upper and lower squares to obtain the six irreps above.
A ligand symmetry orbital can mix with a metal combination only if their irreps match. Since the donor levels lie below the metal levels, mixing produces a lower ligand-dominated bonding orbital and a higher metal-dominated antibonding orbital. In a two-level model with ,The requested metal-dominated levels are shown in the right-hand column; the lower ligand-dominated partners are deliberately outside this diagram.
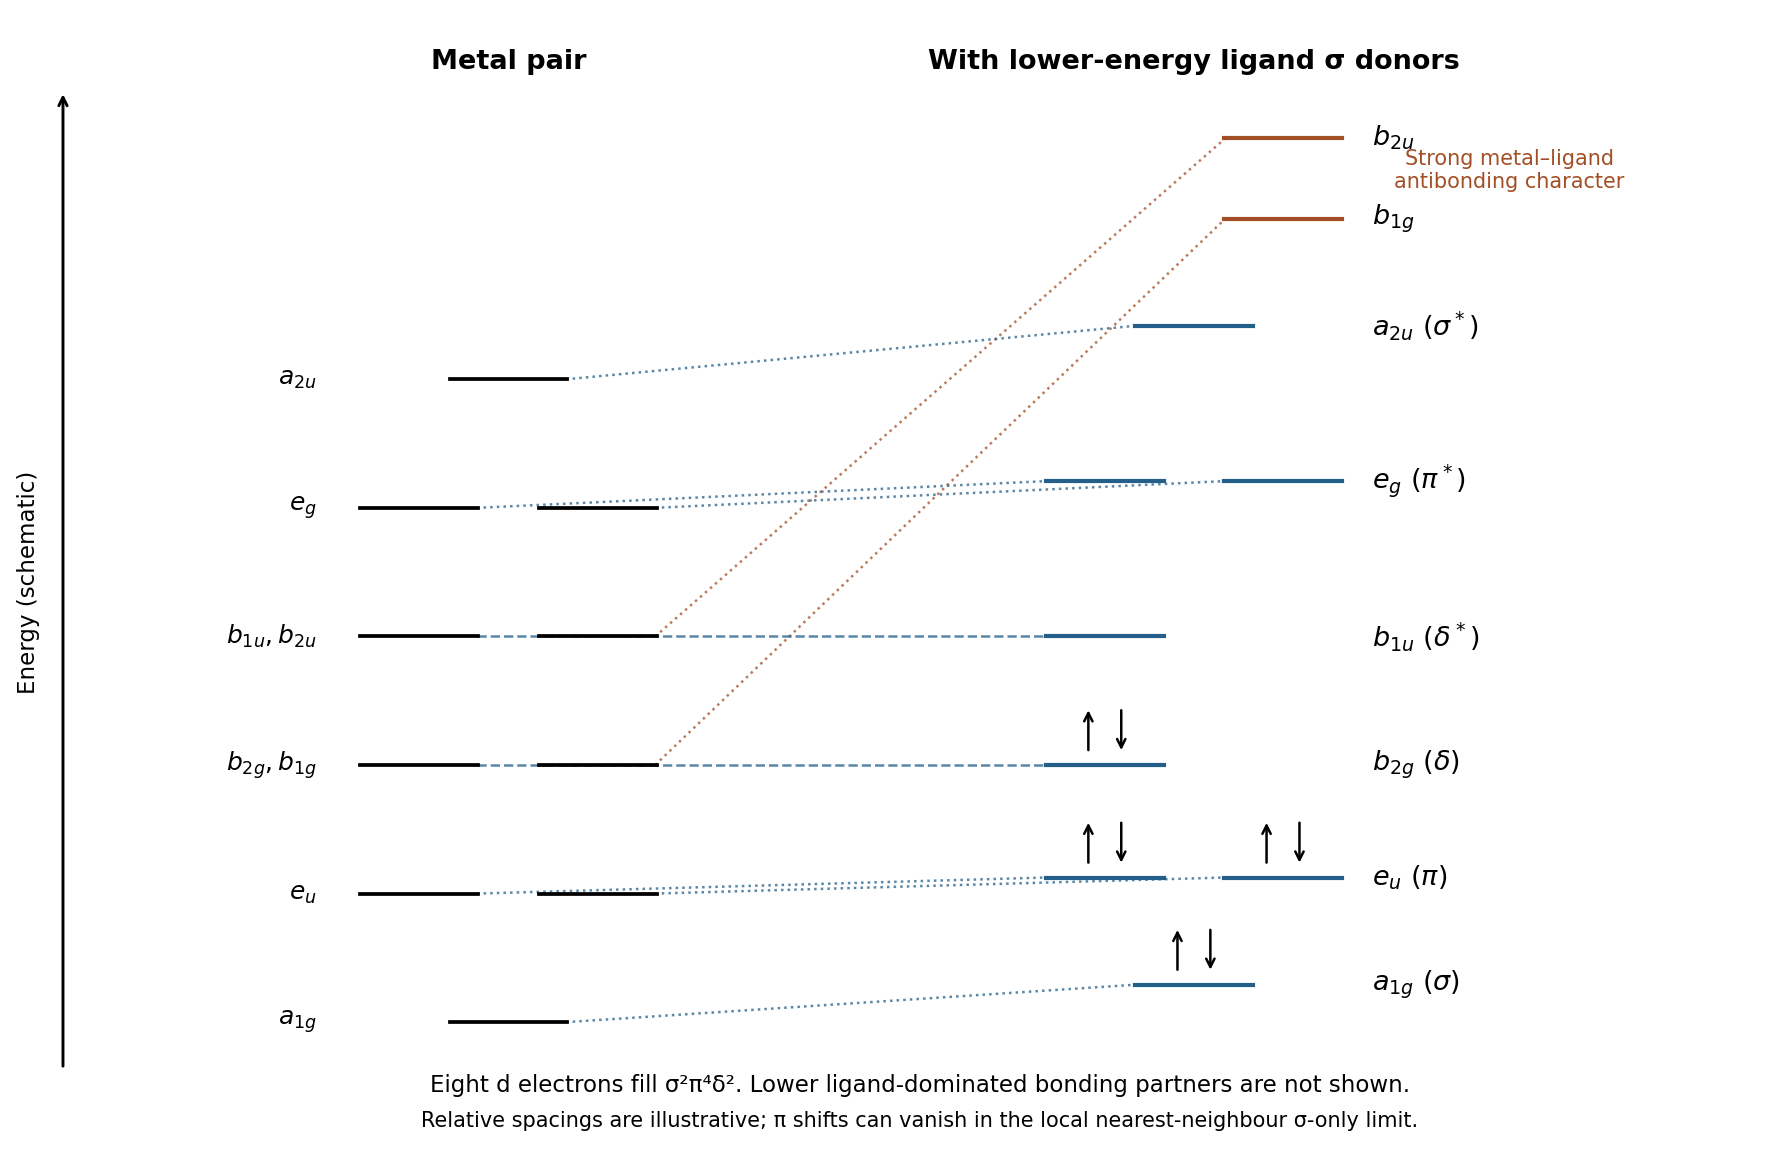


The lobes point directly at the chlorides, so their and combinations are strongly destabilized. The combinations can also interact through their equatorial density. The symmetries have ligand partners, so weaker shifts are symmetry-allowed; their local nearest-neighbour sigma overlap vanishes in the ideal planar geometry, making their shifts much smaller in that approximation. Symmetry allowance alone never sets an overlap integral's size.
There are no ligand sigma combinations of or symmetry, so the -derived delta pair is unaffected by the stipulated sigma-only mixing. In particular the low level survives while the other delta bonding channel becomes strongly metal–ligand antibonding. The plotted spacings and ordering of remote unoccupied levels are schematic; the robust conclusion is the four low Mo–Mo bonding orbitals separated from antibonding or strongly ligand-destabilized orbitals.
The ligand field leaves one low sigma bond orbital, two low pi bond orbitals and one low delta bond orbital. Their closed-shell occupation isIt requires eight metal d electrons, or four per Mo, hence Mo(II). With eight ligands, charge balance givesOxidizing removes an electron from a Mo–Mo bonding orbital; reducing populates a higher orbital with Mo–Mo antibonding or strong metal–ligand antibonding character. Thus the state has the particularly favourable filled bonding configuration in this model. This is an orbital explanation of a preferred charge, not a claim that charge alone determines solution redox stability independently of counterions or solvent.
The eclipsed geometry also follows naturally from the occupied delta bond: it aligns the four-lobed functions. Twisting the chloride squares toward a staggered geometry reduces this overlap and weakens that contribution to the quadruple bond.
Take the eight Mo–Cl bond-length changes as a stretching basis. These scalar coordinates have the same permutation representation as the eight sigma donors:Infrared spectroscopy detects modes transforming as a dipole component: is and is . There are therefore two IR fundamentals, one and one doubly degenerate frequency.
Raman spectroscopy detects modes transforming as polarizability components. The relevant modes are , and , giving three Raman fundamentals, of which is doubly degenerate. The remaining stretch is silent in both ordinary spectra.
Thus the prediction isThe zero follows from infrared and Raman mutual exclusion: the molecule has inversion symmetry, dipole components are ungerade, and polarizability components gerade. This counts resolved fundamental frequencies of the ideal ion, not separate lines for degenerate components, isotope splittings or accidental numerical overlaps between distinct modes.
Articles by others on the same topic
There are currently no matching articles.