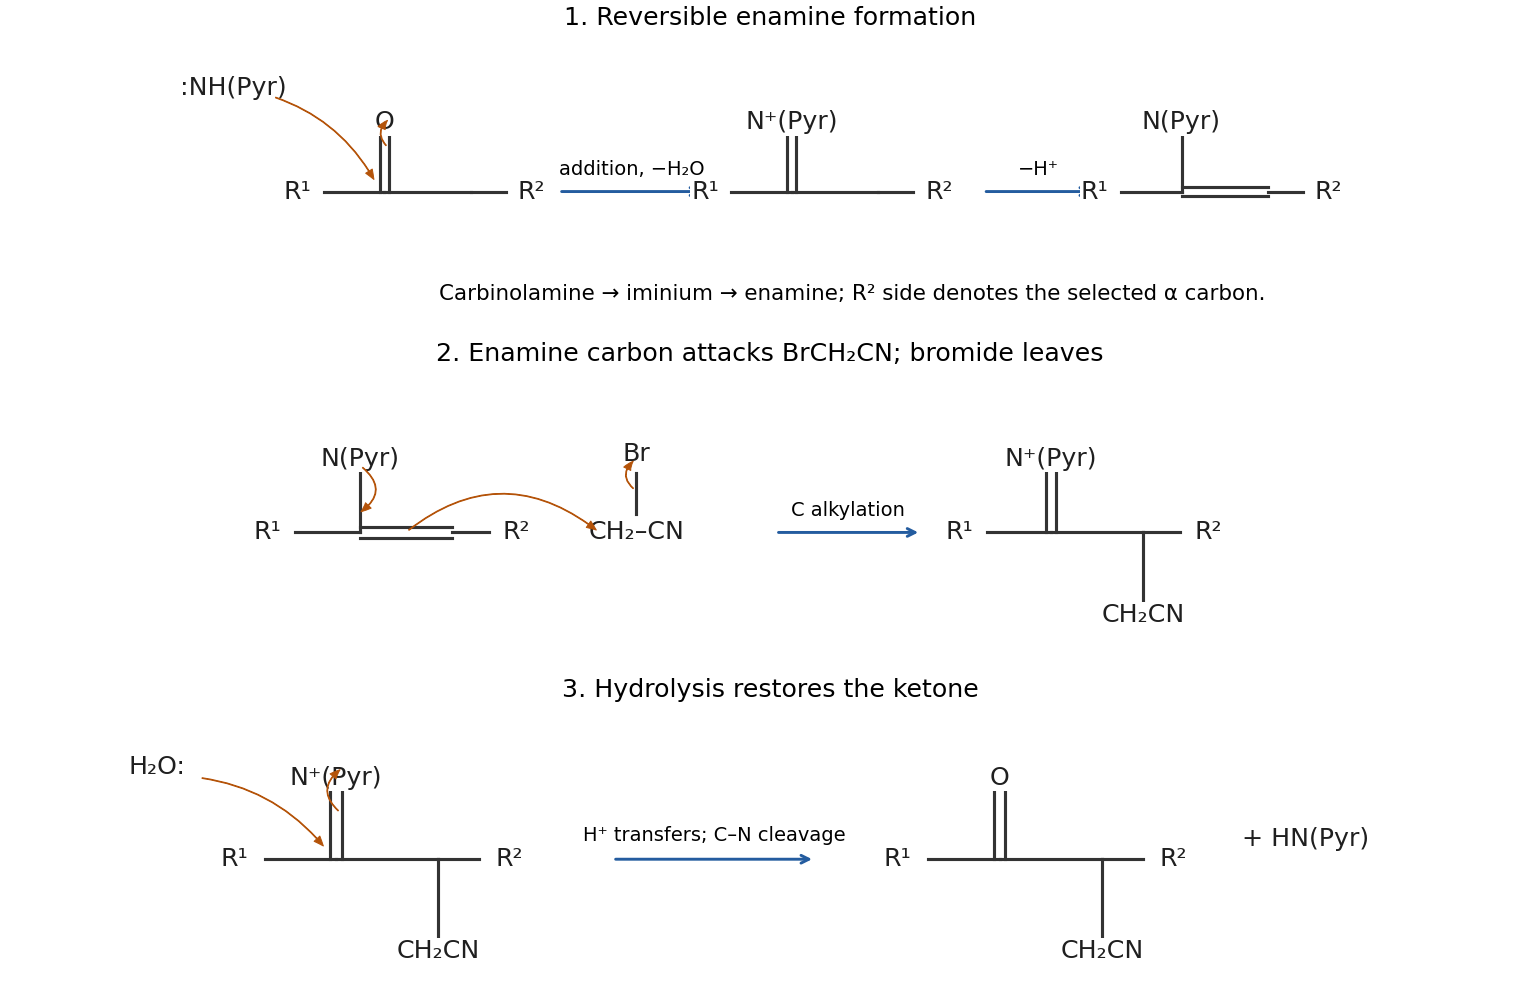
The first stage is a Stork enamine reaction. Pyrrolidine attacks the ketone carbonyl, and proton transfers give a carbinolamine. Loss of water gives an iminium ion; removal of the benzylic alpha proton produces the enamine conjugated with the aromatic ring. Removal of water and prolonged heating favour this condensation. That conjugated enamine is preferred over the alternative with its double bond on the other side of the original carbonyl.
The nucleophilic enamine carbon attacks the methylene carbon of bromoacetonitrile by an SN2 reaction, displacing bromide and making the carbon–carbon bond. It is the carbon beside the aromatic fusion that acquires , as in D. Finally, water adds to the alkylated iminium carbon; proton transfers and cleavage of the C–N bond regenerate the ketone and release pyrrolidine. This explains carbon alkylation, its position, and restoration of the carbonyl rather than treating the amine as a permanent substituent.

Hydride removes the remaining alpha proton at the substituted benzylic carbon, releasing hydrogen and forming the conjugated enolate. Its oxygen attacks the carbonyl carbon of allyl chloroformate. Collapse of the tetrahedral intermediate expels chloride and gives the allyl enol carbonate, with the double bond between the original carbonyl carbon and the cyanomethyl-bearing carbon.
This is O-acylation of an enolate: the carbonyl oxygen is trapped as , retaining the enol double bond. The competing carbon-centred resonance representation explains possible C-acylation in other circumstances, but it would give a different connectivity from E. The indicated product selects the oxygen-trapping pathway and the conjugated enolate regioisomer.
Articles by others on the same topic
There are currently no matching articles.