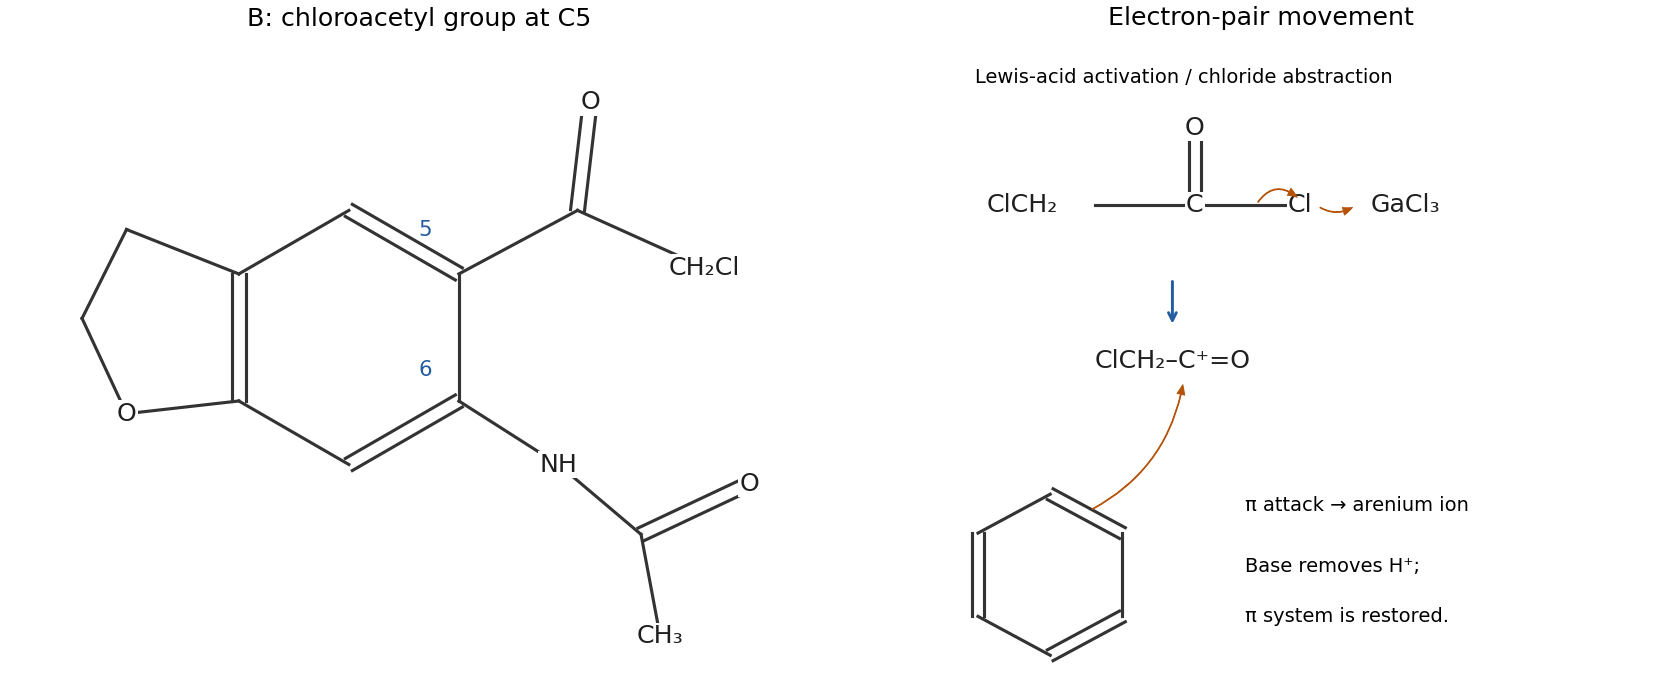
Number the fused heterocycle as a 2,3-dihydrobenzofuran: oxygen is position 1 and the acetamido substituent occupies aromatic position 6. Position 5 is the preferred site of electrophilic substitution. It is para to the ring oxygen and ortho to the acetamido group, so both groups stabilize its Wheland intermediate by resonance donation. Position 7 also receives donation, but lies between the two substituents and is more crowded. Position 4 lacks the same combination of directing effects.
The aryl oxygen donates a lone pair into the aromatic system. The acetamido nitrogen can also donate into the ring, although its lone pair is shared with its own carbonyl group, so it is less activating than a free amino group. Comparing resonance-stabilized intermediates and steric accessibility gives the position-5 prediction; simply counting heteroatoms does not.
The product is N-[5-(2-chloroacetyl)-2,3-dihydrobenzofuran-6-yl]acetamide, with an aromatic substituent at position 5. Substitution of one aromatic hydrogen by gives the stated molecular formula. This positional assignment also agrees with the isolated intermediate reported in the original process study.[ref]
GaCl3 activates chloroacetyl chloride by Lewis-acid coordination and chloride abstraction, represented by the acylium/contact-ion-pair limit . The position-5 aromatic bond attacks the acyl carbon, giving a Wheland intermediate. Removal of the proton at that carbon restores aromaticity. Product–Lewis-acid coordination is reversed on work-up. The carbon-bound chlorine survives; the chlorine on the acyl carbon is the leaving group.
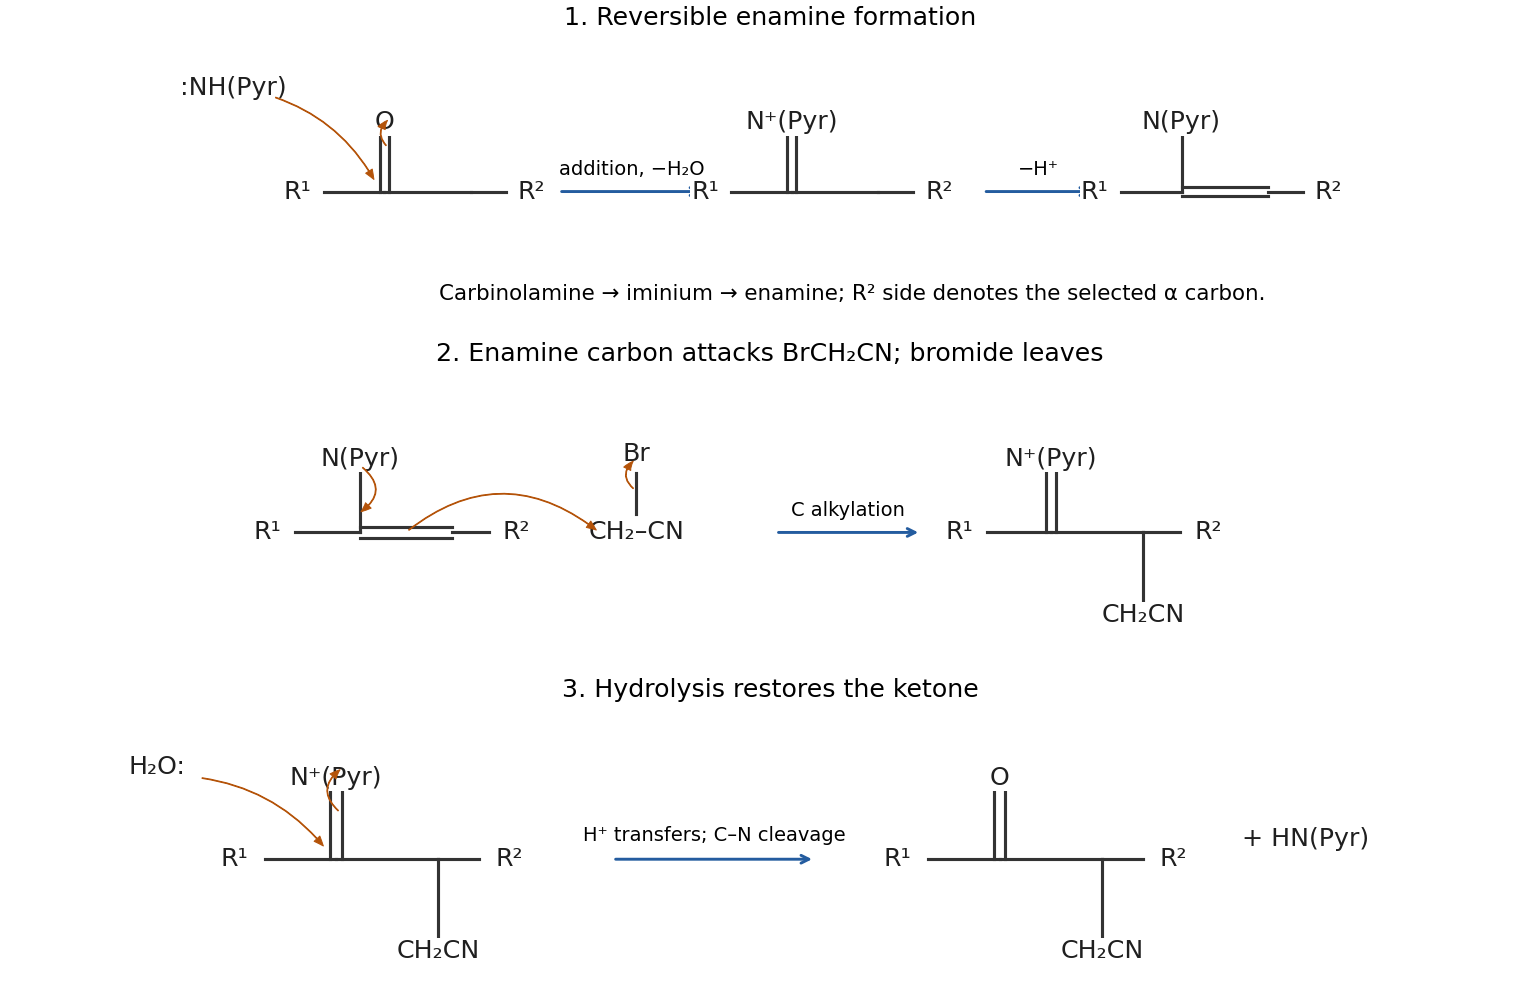
The first stage is a Stork enamine reaction. Pyrrolidine attacks the ketone carbonyl, and proton transfers give a carbinolamine. Loss of water gives an iminium ion; removal of the benzylic alpha proton produces the enamine conjugated with the aromatic ring. Removal of water and prolonged heating favour this condensation. That conjugated enamine is preferred over the alternative with its double bond on the other side of the original carbonyl.
The nucleophilic enamine carbon attacks the methylene carbon of bromoacetonitrile by an SN2 reaction, displacing bromide and making the carbon–carbon bond. It is the carbon beside the aromatic fusion that acquires , as in D. Finally, water adds to the alkylated iminium carbon; proton transfers and cleavage of the C–N bond regenerate the ketone and release pyrrolidine. This explains carbon alkylation, its position, and restoration of the carbonyl rather than treating the amine as a permanent substituent.
Hydride removes the remaining alpha proton at the substituted benzylic carbon, releasing hydrogen and forming the conjugated enolate. Its oxygen attacks the carbonyl carbon of allyl chloroformate. Collapse of the tetrahedral intermediate expels chloride and gives the allyl enol carbonate, with the double bond between the original carbonyl carbon and the cyanomethyl-bearing carbon.
This is O-acylation of an enolate: the carbonyl oxygen is trapped as , retaining the enol double bond. The competing carbon-centred resonance representation explains possible C-acylation in other circumstances, but it would give a different connectivity from E. The indicated product selects the oxygen-trapping pathway and the conjugated enolate regioisomer.
Pyrrolidine and 2-methylcyclohexanone first form an enamine, through carbonyl addition, carbinolamine formation, dehydration to an iminium ion, and alpha deprotonation. The less hindered enamine carbon is on the side opposite the methyl-bearing alpha carbon. It attacks the terminal beta carbon of methyl vinyl ketone, while the conjugated system shifts to oxygen. Protonation gives an alkylated iminium species; aqueous acid then hydrolyses it back to the ketone.
The result is a Michael addition product: the previously unsubstituted alpha carbon bears . In drawing the arrows, start the new C–C bond at the enamine carbon, not at nitrogen. The iminium bond is restored during addition and removed by hydrolysis. The sequence is an enamine-mediated conjugate addition, rather than direct addition to the methyl vinyl ketone carbonyl.
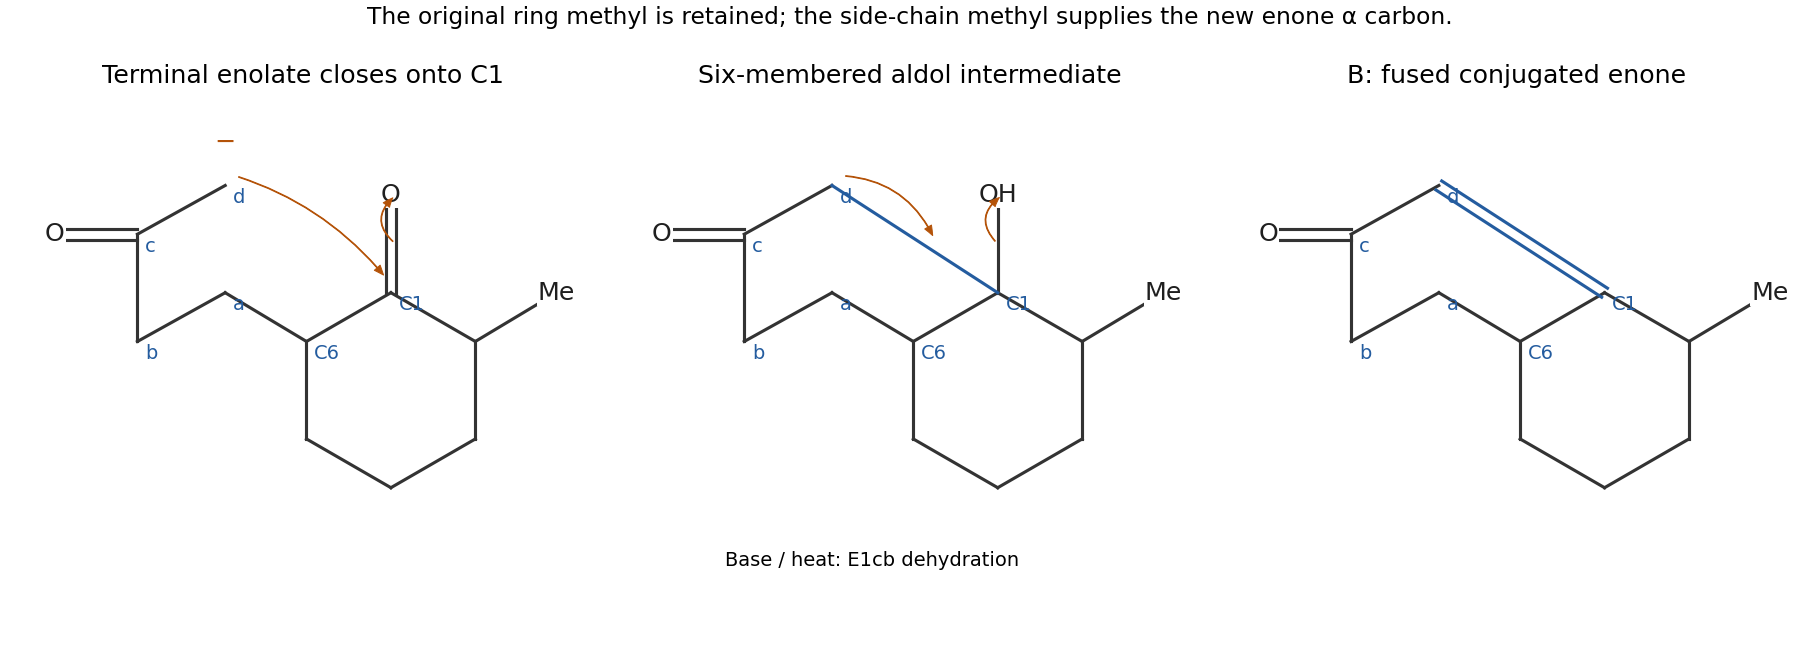
A undergoes a Robinson annulation. For an unambiguous connectivity description, call the original ring carbonyl carbon C1 and the side-chain-bearing alpha carbon C6. Label the appended chain , where d is initially methyl. Base makes the enolate at d, which attacks C1 intramolecularly to close the six-membered ring . The carbonyl electrons move to oxygen, and protonation gives the beta-hydroxy ketone.
An E1cb elimination then removes the proton on d and the hydroxy group on C1, giving the conjugated enone with C1=d and a carbonyl at c. The original ring methyl remains on the carbon next to C1 on the other side. The two rings share the C1–C6 bond. The molecular formula is consistent with loss of one water molecule from the diketone.
The aniline nitrogen attacks the ipso carbon bearing fluorine. The aromatic electrons shift to give a Meisenheimer complex, with negative charge delocalized to the positions ortho and para to the attacked carbon. At either nitro substituent, that charge can be delocalized further onto nitro oxygen. Subsequent proton transfer and fluoride elimination restore aromaticity and give the diarylamine.
Both the ortho and para nitro groups stabilize the addition intermediate in 1-fluoro-2,4-dinitrobenzene. The para-mononitro compound lacks the additional ortho stabilization, so addition is less favourable and generally slower under the stated conditions. This comparison concerns stabilization of the addition pathway; the presence of fluorine does not make the reaction an ordinary aliphatic SN2 reaction.
The qualitative nucleophilicity order is -OMe -Me H -Cl -NO, matching the product-yield order at the stated stopping time. Electron donation raises the availability of the aniline lone pair for attack. A para methoxy group donates by resonance, while methyl is a weaker donor. Chlorine withdraws inductively and gives an aniline less nucleophilic than the unsubstituted compound. A para nitro group strongly withdraws by resonance: electron flow from nitrogen through the ring into the nitro group stabilizes a charge-separated form and greatly reduces nucleophilicity.
The key arrows for the nitroaniline comparison move the nitrogen lone pair into its own aromatic ring, shift the ring bonds toward the para nitro group, and shift a nitro N=O bond toward oxygen. The methoxy donor produces the opposite electronic bias. These are finite-time yields, so they support a kinetic explanation qualitatively; they are not by themselves measured rate constants or equilibrium constants.
Base deprotonates the sulfonamide. Its nitrogen attacks one epoxide carbon from the back, breaking the C–O bond at that carbon and producing an adjacent alkoxide. The nitrogen and oxygen attachments on the cyclopentane are therefore trans. The alkoxide then attacks the aromatic carbon bearing the fluorine ortho to the sulfonyl group. Intramolecular nucleophilic aromatic substitution expels fluoride and closes a seven-membered heterocycle with the path . The para fluorine remains.
This tandem epoxide-opening/ring-closing pathway is required by the formula of F: simple epoxide addition would retain both fluorines, whereas F contains only one. In the second stage, pyrrolidine replaces the remaining para fluorine by another nucleophilic aromatic substitution, producing G. The mass balance is F plus pyrrolidine minus HF, as required by the given formula of G.
The meso epoxide has enantiotopic ends. With achiral reagents, opening at either end gives an enantiomeric product, so F is a racemic pair with the same trans relative configuration; G retains that configuration.
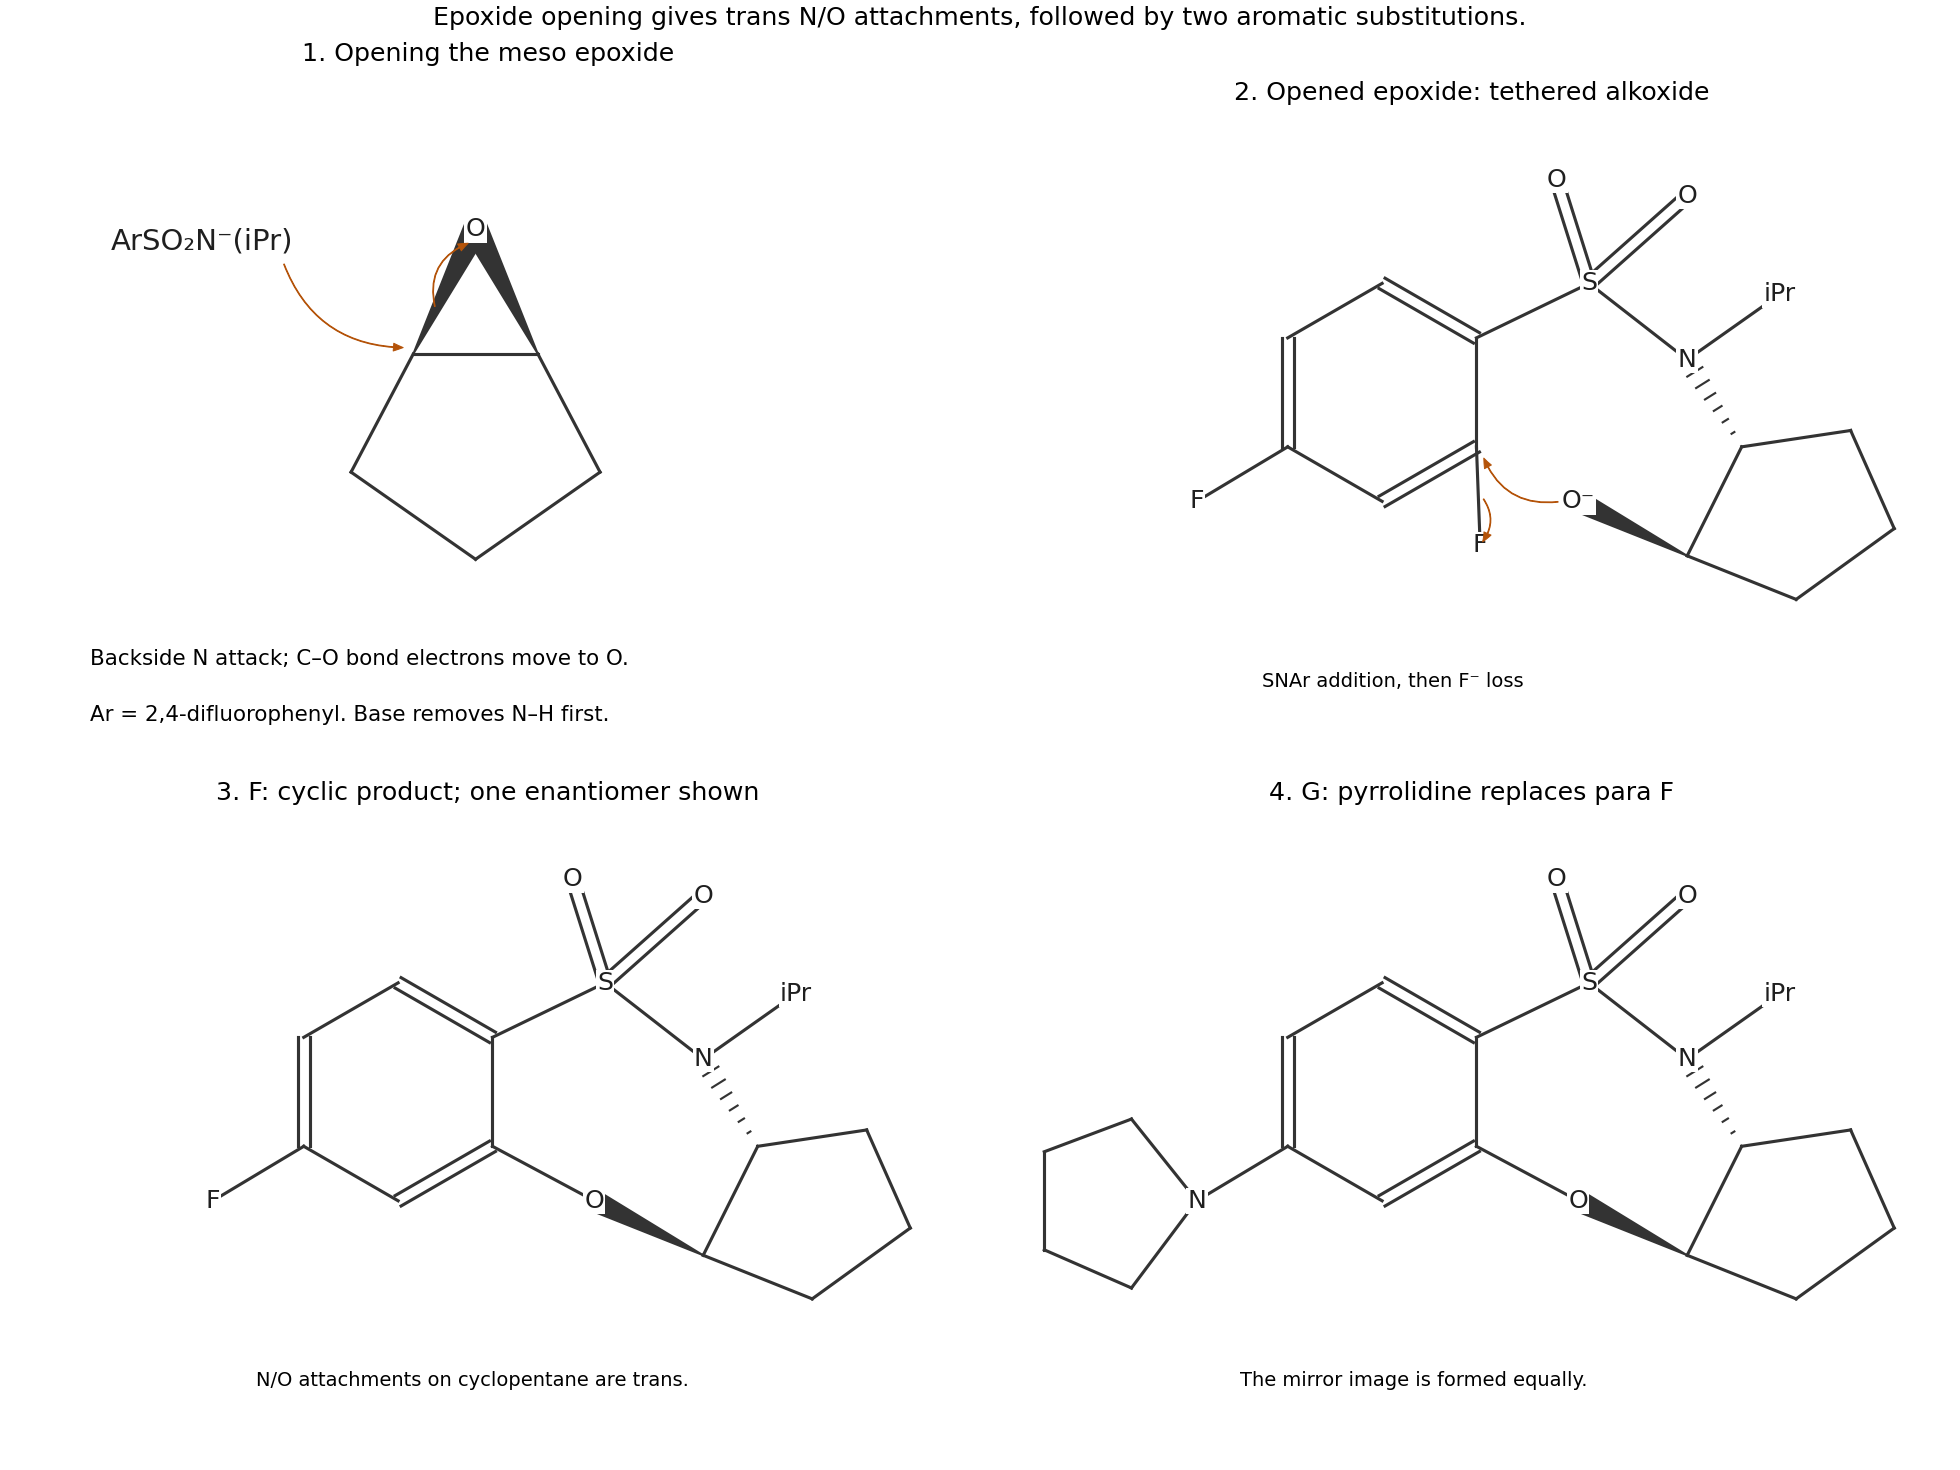
Trans epoxide-opening cascade, seven-membered ring closure, and replacement of the remaining aryl fluoride
. Details. The relevant quantity is the H–C–C–H dihedral angle, through the Karplus relation, rather than the labels cis and trans alone. Couplings tend to be larger near eclipsed or antiperiplanar arrangements and smaller nearer a right angle. The observed value averages the accessible conformations.
In A, the saturated five-membered ring is puckered. A preferred envelope-like conformation can place the trans hydrogens closer to an antiperiplanar arrangement, giving a slightly larger coupling than the cis pair. In B, the double bond restricts ring geometry toward a flatter arrangement: the cis hydrogens are more nearly eclipsed, whereas the trans pair has an oblique dihedral angle. The cis coupling is then larger. Approximate Newman projections illustrate these trends; exact angles cannot be recovered from four coupling constants without a calibrated relation and conformational populations.
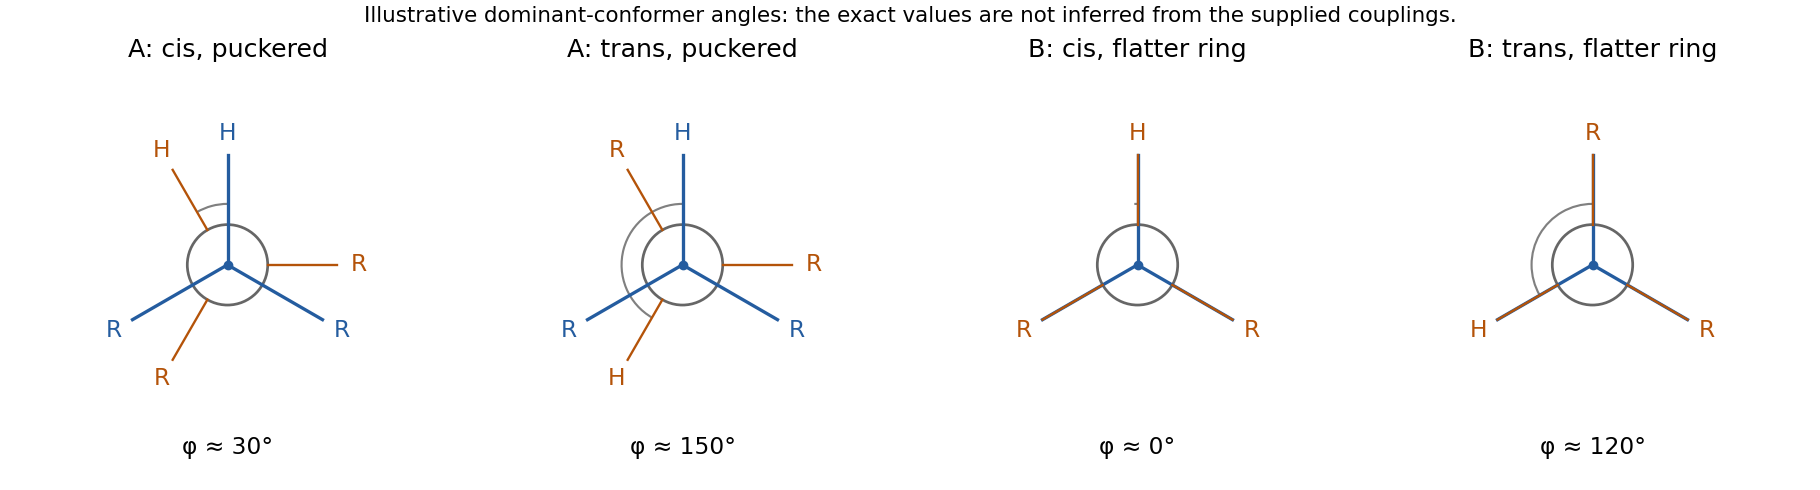
Illustrative Newman projections explaining why cis/trans coupling order depends on five-membered ring geometry
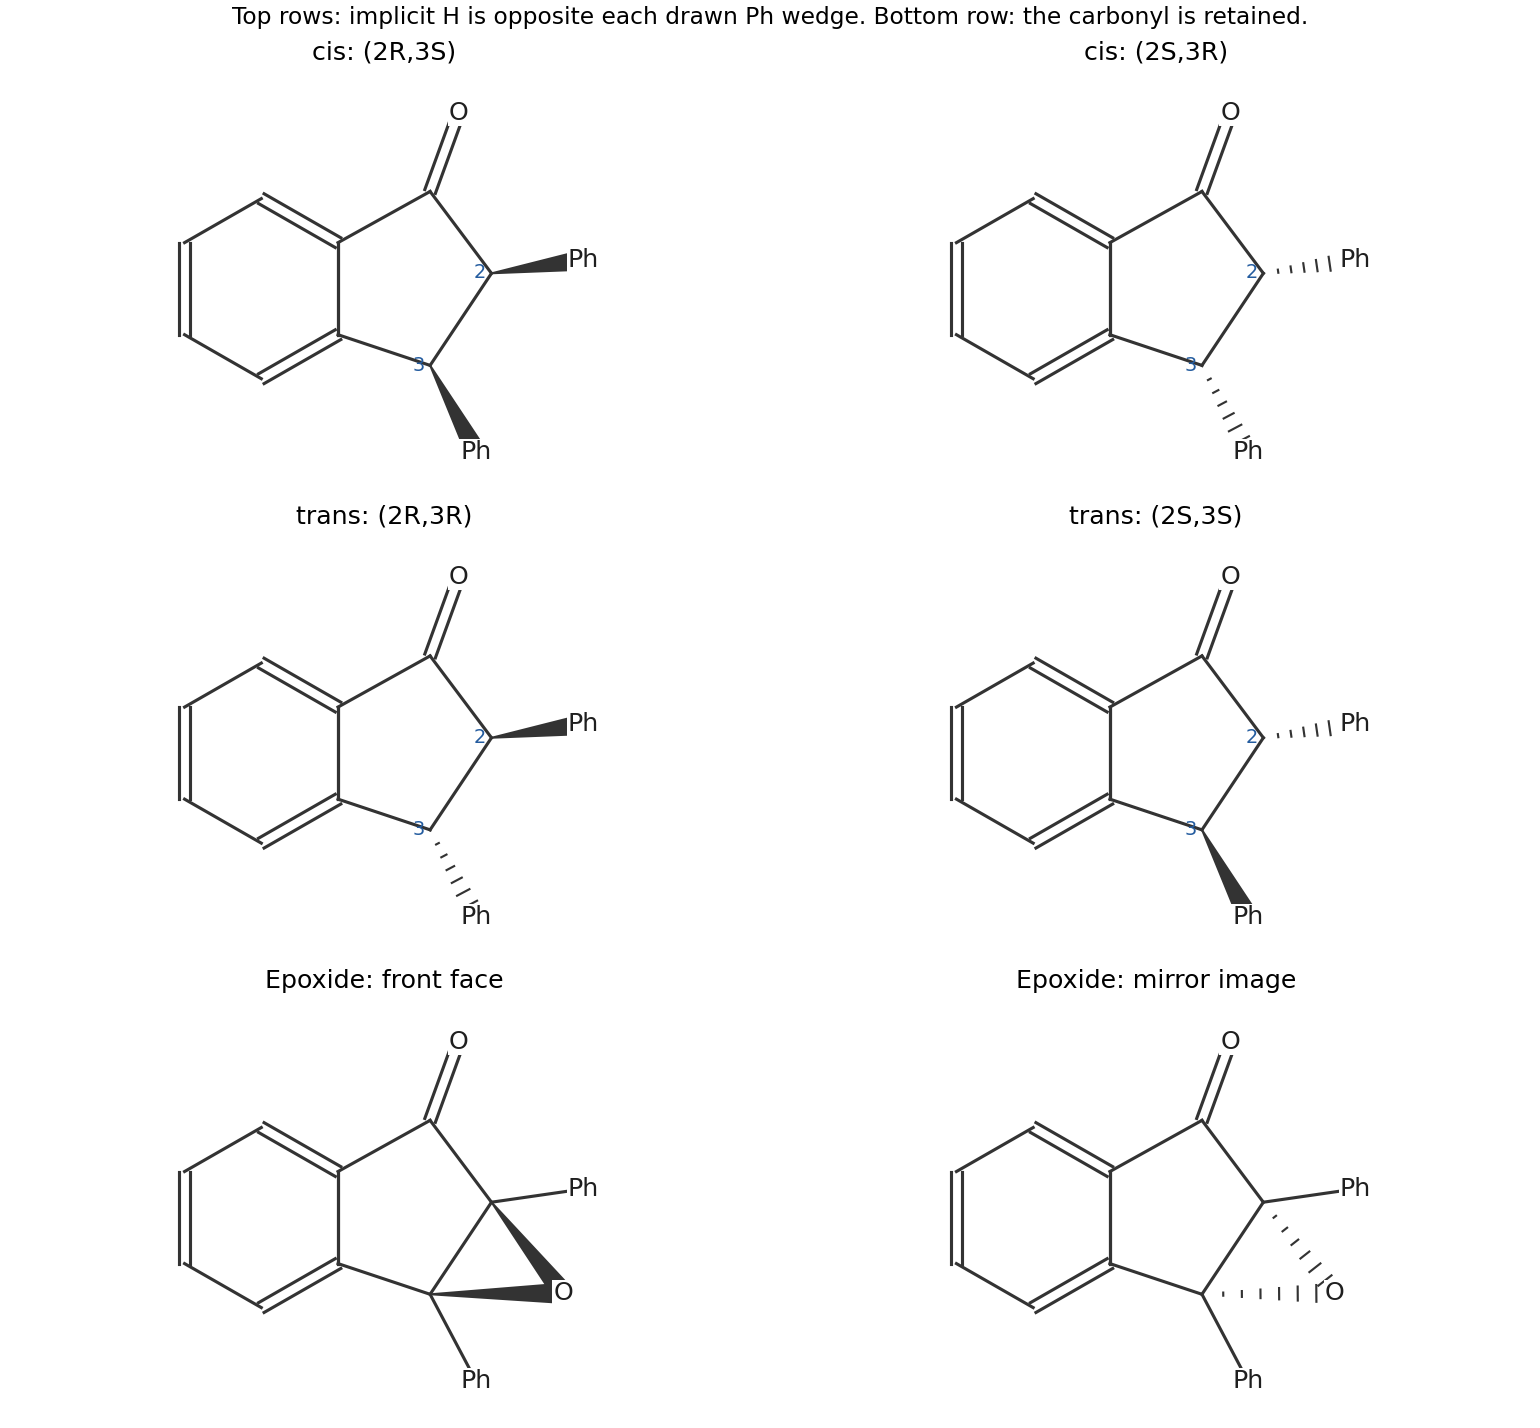
. Details. Reduction makes stereogenic centres at C2 and C3 of the indan-1-one framework. The four products are the cis pair and , and the trans pair and . Each pair consists of enantiomers; any cis member is diastereomeric to either trans member. There is no meso product because the carbonyl-containing and aromatic sides of the ring do not make C2 and C3 symmetry-equivalent.
For checking the Cahn-Ingold-Prelog priority rules, at C2 the order is carbonyl-side carbon phenyl C3 H. At C3 it is fused aromatic carbon phenyl C2 H. The fused aromatic path outranks the separate phenyl path because it reaches the carbonyl-containing part of the framework.
At 400 MHz, a separation in ppm corresponds to Hz. The two multiplets near 5.10 and 4.47 ppm each have a quoted separation of 0.02 ppm, giving Hz. The multiplets near 4.625 and 3.865 ppm each have separation 0.01 ppm, giving Hz. These are approximate values from the rounded peak labels.
The fused unsaturated/aromatic framework restricts the five-membered ring, as in the comparison in (a). Assign the larger coupling to cis and the smaller to trans. Matching equal couplings and integrals pairs the 5.10/4.47 signals with one another, and the 4.625/3.865 signals with one another. The integral ratio isWith an achiral substrate and achiral catalytic environment, each diastereomer is expected as a racemic enantiomeric pair: cis and trans . The reported spectrum requires both diastereomers; the reagent label alone cannot override that evidence. Their ratio does not establish whether the selectivity arose in the initial surface addition or after product epimerization.
Only the low-coupling pair remains, so the detected product is the trans diastereomer, with approximate coupling 4 Hz. The reaction is highly diastereoselective within the spectrum's detection limit. Its possible absolute configurations are and .
A conventional proton NMR spectrum in an achiral environment cannot distinguish those enantiomers: they have the same shifts and couplings. Thus a chiral catalyst plus one set of resonances does not establish an enantiomeric excess or an absolute configuration. Chiral chromatography, an appropriate chiral NMR method, or a calibrated stereochemical comparison would be needed.
The products are the two mirror-image 2,3-epoxy-2,3-diphenylindan-1-ones, shown in the bottom row of the stereoisomer diagram. The original carbonyl remains, and an additional oxygen bridges the former double bond, giving the stated formula.
In the Weitz-Scheffer epoxidation, hydroperoxide anion adds conjugatively to the enone. The resulting enolate closes onto the attached peroxide oxygen with O–O bond cleavage and loss of hydroxide. Attack on the two faces of the achiral planar substrate gives the two enantiomers in equal amounts. This is a racemic epoxide pair, rather than a one-to-one mixture of different diastereomeric epoxide connectivities.
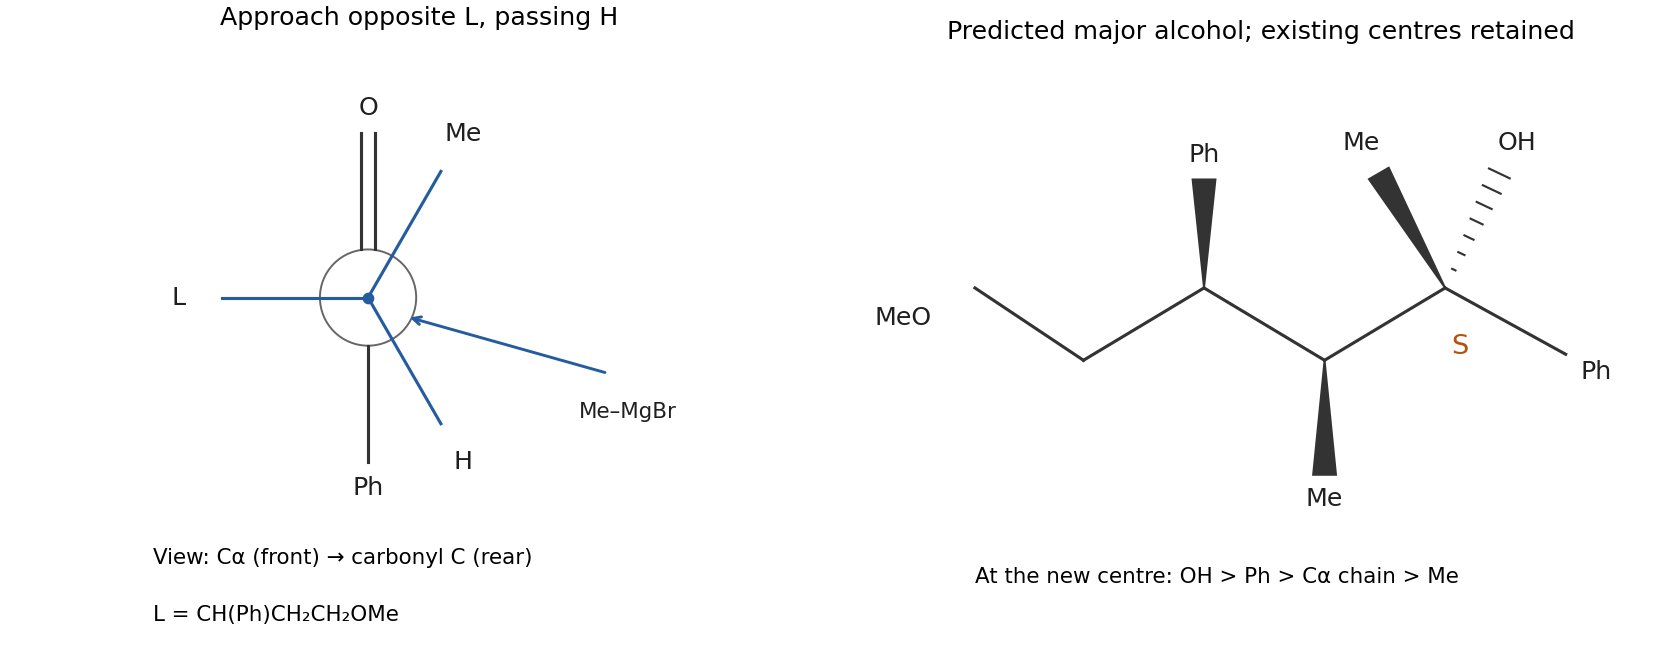
Apply the Felkin-Anh model to the stereocentre alpha to the ketone. Its large group is the chain continuing through the phenyl-bearing carbon, its medium group is methyl, and its small group is hydrogen. In the preferred transition-state arrangement, put the large group approximately perpendicular to C=O and approach from the opposite face, passing the small hydrogen along a Bürgi-Dunitz trajectory. The remote ether cannot form the usual small alpha-alkoxy chelate, and the coordinating solvent further supports using an open-addition prediction.[ref]
Felkin-Anh and Cram Chelate by University of Windsor. pp. 1–2: perpendicular large group and nucleophilic approach past the small group
The predicted new stereocentre is . In the question's zigzag projection, this corresponds to the new OH bond directed behind the page and the newly introduced methyl toward the reader, with the original stereocentres unchanged. The new-centre priority order is OH phenyl the substituted alpha-carbon chain methyl. This is a model-based major-product prediction, rather than a quantitative diastereomeric ratio.
Protonation of a spiroacetal oxygen makes the adjacent C–O bond easier to cleave. Reversible opening gives an oxocarbenium ion, in which the spiro carbon is locally planar. The tethered alcohol can attack either face; deprotonation closes the acetal and regenerates acid. Repetition interconverts the spiro epimers while retaining the other stereogenic centres. Methanol can participate in reversible proton transfer or trapping, but there is no need to invert the remote carbon centres.
The diagnostic feature is the large vicinal coupling characteristic of an axial–axial relationship in a chair. The td patterns for and contain two large couplings to neighbouring axial hydrogens, so both are axial. The ddd patterns for and also include a large ring vicinal coupling, with the smaller values reflecting the other ring hydrogen and the side chain. The assignments areThe chemical shifts alone do not determine this assignment. In particular, three unequal splittings of a methine are not a geminal-coupling pattern.
Use the upper-ring chair placing or axial. The aryl ester substituent on that carbon is then equatorial. Use a lower-ring chair placing or axial, with the attached side chain equatorial. The spiro configuration distinguishes the two structures: in A, the lower-ring oxygen bond is axial with respect to the upper ring, while the upper-ring oxygen bond is equatorial with respect to the lower ring. In B, those roles reverse.
Thus the oxygen placements at the shared spiro carbon are axial/equatorial for A and equatorial/axial for B. Side-chain conformations are unnecessary, but changing the lower chair without tracking its substituent positions would change this conclusion.
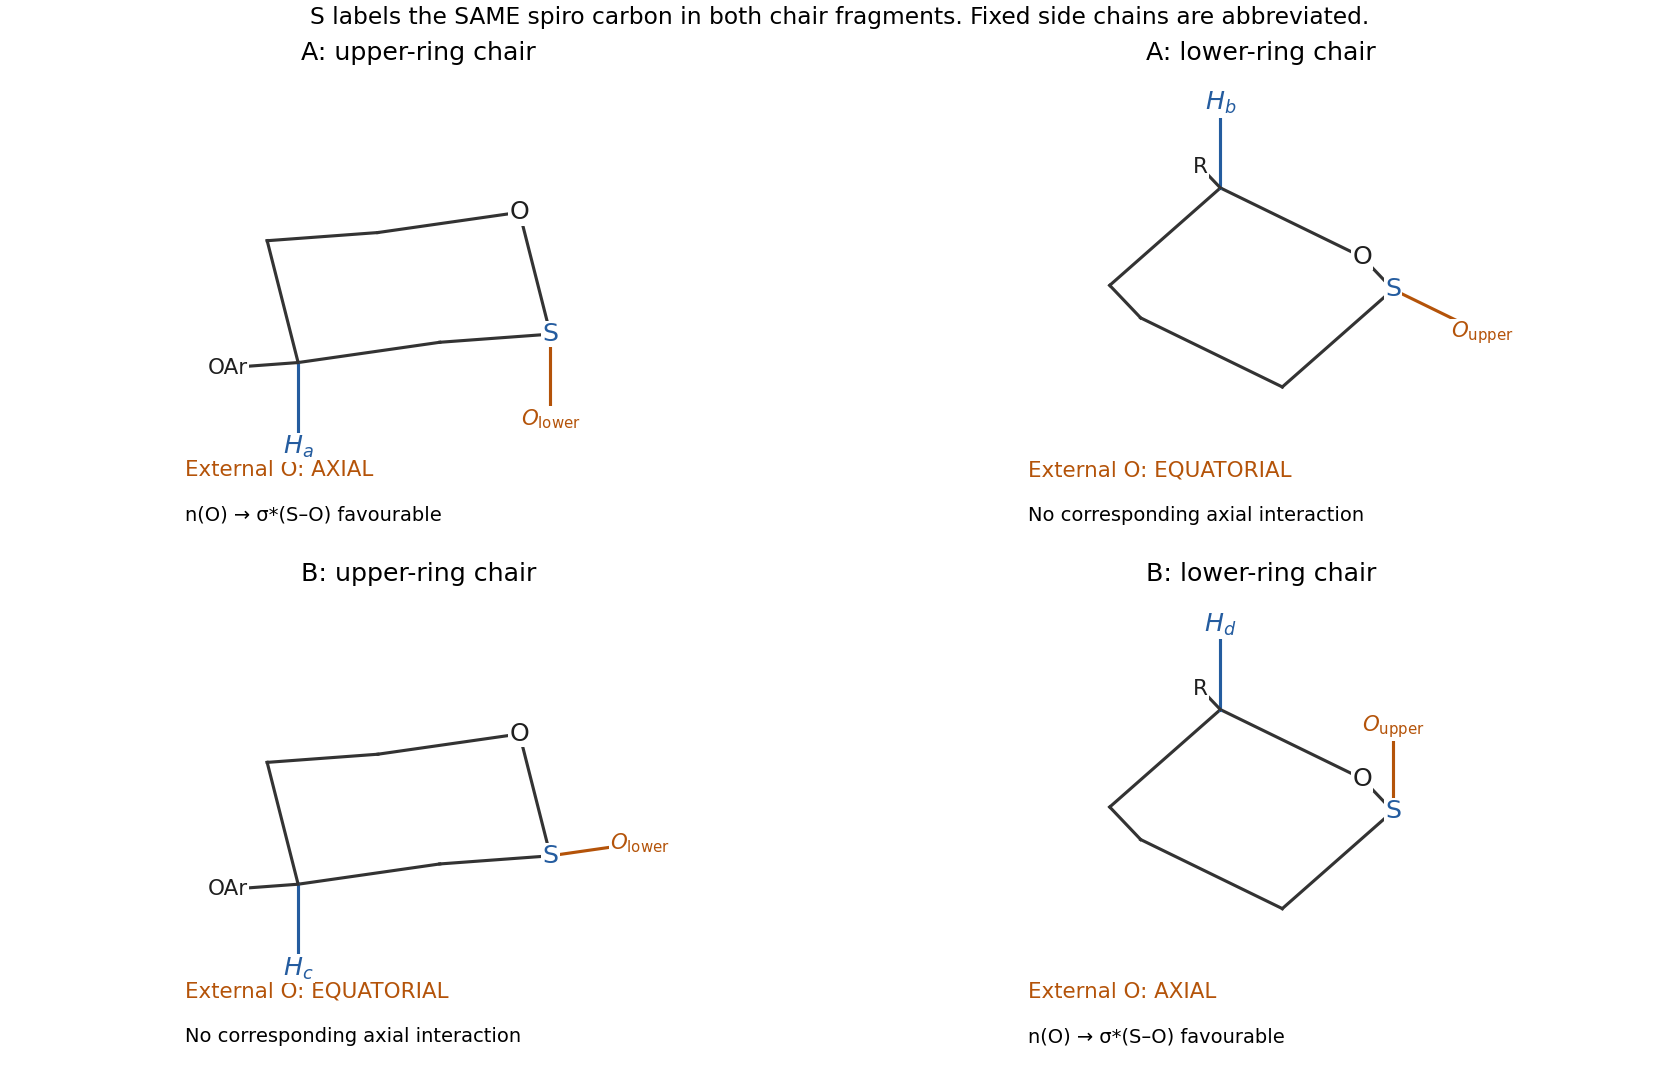
Chair assignments for the two spiroacetals: axial reporter hydrogens and one axial exocyclic oxygen in each isomer
. Details. The anomeric effect is the preference, in suitable cyclic acetals, for an electronegative substituent at an anomeric carbon to occupy an axial position even when simple steric reasoning favours equatorial. An important orbital contribution is donation from an endocyclic oxygen lone pair into the antibonding orbital of the adjacent exocyclic C–O bond, . The chair's axial arrangement supplies favourable antiperiplanar overlap. Electrostatic and solvation contributions can also affect the observed conformational preference.
Each isomer has one favourable anomeric interaction. A has the favourable axial exocyclic oxygen in the upper-ring chair; B has it in the lower-ring chair. The NMR-constrained conformations therefore do not provide a doubly anomeric versus non-anomeric comparison.
Both can keep important carbon substituents equatorial and each obtains one anomeric stabilization. Appreciable amounts of both are therefore a reasonable prediction, rather than a strong preference based solely on counting anomeric interactions. The environments of the two ring oxygens are not identical, so this argument does not determine an exact one-to-one equilibrium ratio; quantitative preference would require further energetic information.
For hydrogen-bond acceptance, the three strongest groups are the terminal carboxylate, the Cys–CO–N(Pro) tertiary-amide carbonyl, and the Pro–CO–NH(Tyr) secondary-amide carbonyl, in that qualitative order. Negative charge makes carboxylate a particularly strong acceptor. Both amide oxygens accept strongly; greater nitrogen alkyl donation generally makes a tertiary amide oxygen more basic than the comparable secondary-amide oxygen. This last ordering is a functional-group estimate, not a precise calculation of a folded tripeptide's site parameters.
For donation, the order is terminal ammonium tyrosine phenolic OH the peptide NH between Pro and Tyr. The thiol is a weaker donor. Protonated ammonium has no available lone pair to accept; the proline amide nitrogen has no N–H bond to donate, and amide nitrogen lone pairs are delocalized into carbonyl groups. The carboxylate has no O–H donor in the drawn protonation state.
The functional group interaction profile compares donor strength and acceptor strength . In water, strong–strong contacts compensate for desolvation, giving the upper-right black region. Strong–weak contacts lose favourable solvent interactions without adequate replacement, giving the two grey regions. Weak–weak association gives the lower-left black region through the hydrophobic effect.
Ethanol's ethyl groups preferentially solvate weakly polar sites, removing this incentive for their association: the lower-left region becomes approximately neutral. Its hydroxyl group retains competition for hydrogen bonds elsewhere in the profile.[ref]
There are three natural states: an unfolded single strand, a monomeric DNA hairpin, and an intermolecular self-duplex. A useful parsing of the sequence is . The two five-base arms are reverse complements. They can form a hairpin with a five-base-pair stem, a four-base CAAC loop, and a dangling 5′ T.
Two copies can instead pair their arms intermolecularly. The self-duplex has two five-base-pair regions, a central noncomplementary internal region, and dangling T residues; it is not a perfectly paired 15-base-pair duplex. Low temperature favours pairing. At relatively low strand concentration, the unimolecular hairpin avoids the association-entropy cost of a dimer; at sufficiently high concentration, intermolecular pairing becomes more favourable. At high temperature, the unfolded state predominates. Actual crossover temperatures require salt conditions and sequence thermodynamics.
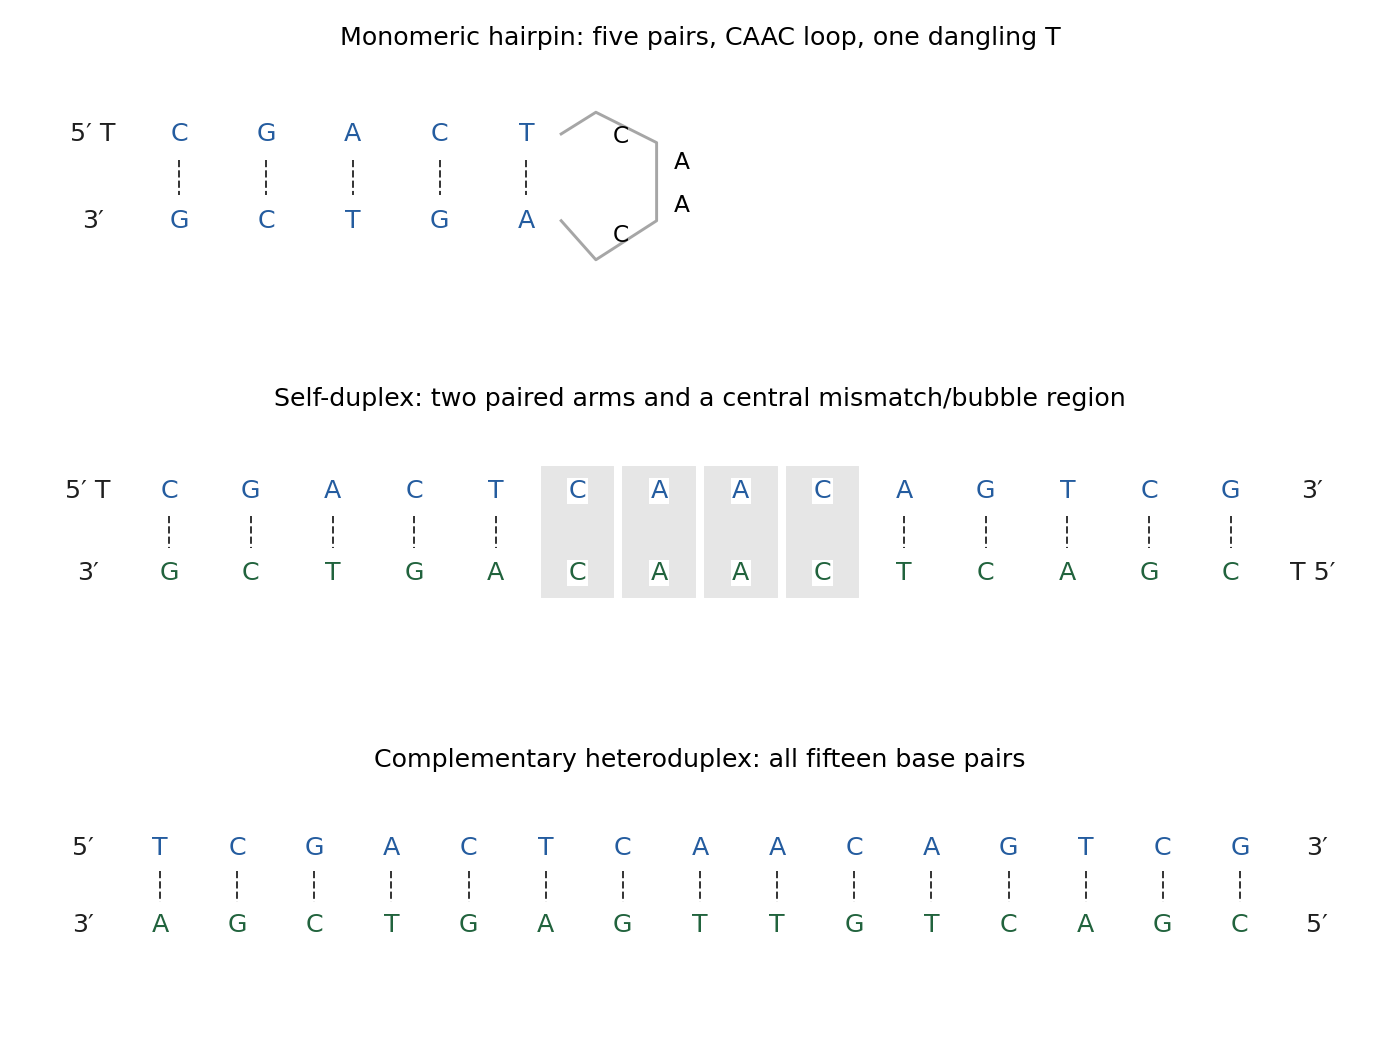
The second strand is the complete reverse complement of the first. They can therefore form a fully matched antiparallel 15-base-pair heteroduplex, replacing the less fully paired hairpin/self-duplex states under conditions supporting hybridization:The lower line is written antiparallel; read it from its own 5′ end to recover the added oligomer.
- : a very small four-membered hydrogen-bonded loop, incompatible with a favourable repeating peptide-backbone geometry.
- : a seven-membered loop associated with a gamma turn; a long regular two-residue-per-turn structure is strongly constrained.
- : the ten-membered hydrogen-bonded rings of a 3-10 helix, with about three residues per turn.
- : the thirteen-membered rings of an alpha helix, with about 3.6 residues per turn.
- : the sixteen-membered rings of a pi helix, with about 4.4 residues per turn.
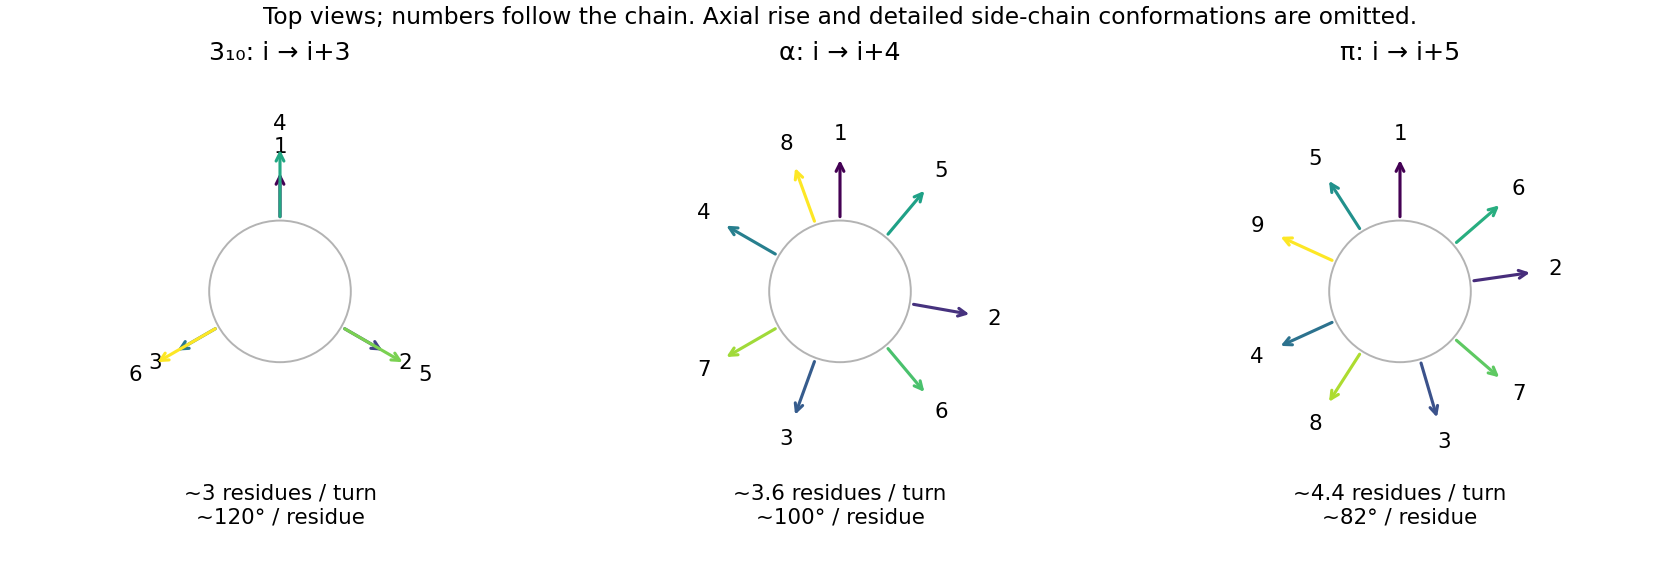


The alpha helix provides a favourable combination of allowed backbone torsions, reasonably linear hydrogen bonds, and side-chain packing. Its successive side chains advance by about , rather than lining up in the three columns of the narrower 3-10 helix. Smaller repeating rings impose more severe backbone constraints; the wider pi helix gives a different, generally less favourable packing arrangement. Short 3-10 and pi-helical segments can still occur in proteins, so this is a preference for a common extended secondary structure, not a prohibition of the alternatives. Proline and sequence-specific interactions can disrupt the idealized backbone donor pattern.
Articles by others on the same topic
There are currently no matching articles.