The central carbon is approximately hybridized, with two opposite sigma bonds and no carbon lone pair; the electron-rich donor ends are sulfur and nitrogen. The 16 valence electrons allow the principal resonance structuresEach has octets and a linear S–C–N skeleton. Carbon's two perpendicular p orbitals support the pi interactions. The first contributor emphasizes the C≡N bond and sulfur-centred negative charge; the second distributes pi bonding over S=C=N and puts negative charge on N. The real anion has partial multiple-bond character and delocalized charge rather than alternating between these drawings.
Nitrogen's greater Allen electronegativity favours electron accumulation there, while sulfur supports a diffuse, polarizable electron-rich region. Thus both terminal atoms are important electron-rich sites relative to central carbon. “Greatest electron density” should not be interpreted as a literal comparison of all core electrons on different elements; the chemically relevant comparison is valence charge, donor density or electrostatic potential. The exact partition of negative charge between S and N is not uniquely determined by electronegativities or Lewis structures alone.
The Hg–S interaction is expected to be stronger than Hg–N by the hard and soft acids and bases argument. is a soft, readily polarizable Lewis acid. The sulfur end of thiocyanate is a softer and more polarizable donor than its nitrogen end, so it better matches mercury's electronic response and covalent bonding preference.
Nitrogen being more electronegative does not by itself make it the preferred donor to a soft metal. Charge concentration and polarizability are distinct properties. The extended solid can still contain Hg–N contacts that complete the bridging network; the prediction concerns their relative bonding preference, not the absence of N coordination.[ref]
Hard and Soft Acids and Bases by Ralph G. Pearson. 1963, primary classification and matching principle
The ordering reflects different connectivities, bond strengths and charge distributions, rather than different elemental composition. Relative to thiocyanate, the supplied formation enthalpies are higher by and , respectively.
For , carbon can make strong C–N multiple bonding with good 2p–2p overlap while keeping its octet. The contributors and avoid a compulsory positive central atom and place the excess negative charge on terminal heteroatoms.
For , an octet description with four bond-order units forces the central nitrogen to formal charge . A representative contributor is , with both charge separation and negative charge on less electronegative carbon. The alternative does not remove that disadvantage. Connectivity prevents simply transferring the favourable thiocyanate charge pattern to this isomer.
For , a central-sulfur octet with four bond-order units gives sulfur formal charge . One illustrative contributor is , which has severe charge separation. Attempts to avoid it by drawing strong expanded-octet multiple bonding at sulfur do not supply the strong, well-matched second-row pi overlap available in C–N bonds. Sulfur 3p functions are larger and less well matched to 2p functions, and high-energy 3d orbitals are not a remedy for this deficiency.
These arguments rationalize the large differences but do not quantitatively predict the supplied enthalpies from electronegativity alone. Real charge delocalization and geometry relaxation matter; formal charges are bookkeeping, not measured atomic charges.
Balancing Hg first, then sulfur, carbon and nitrogen givesThere are two Hg, four S, four C and four N atoms on each side. This is the stipulated initial decomposition, not a claim that further heating cannot change its products.
HgS and are semiconductors; is a wider-gap, insulating material under the stated conditions. None is a metallic conductor. Their finite electronvolt-scale gaps and very low measured conductivities exclude a partially filled, highly conducting metallic band picture.
The insulator/semiconductor boundary is conventional rather than a sharp universal gap cutoff. Calling the material a wide-bandgap semiconductor is also physically reasonable; the table alone does not establish a unique naming boundary. Conductivity is , so its ordering need not follow gap size alone: carrier density, defects, trapping and mobility also matter.
One possible idealized sheet is a network of alternating C/N triazine rings joined through three-coordinate nitrogen bridges. Every carbon has three sigma neighbours and participates in a conjugated pi framework. Each ring nitrogen has two ring neighbours and a lone pair; each bridging nitrogen joins three ring carbons.
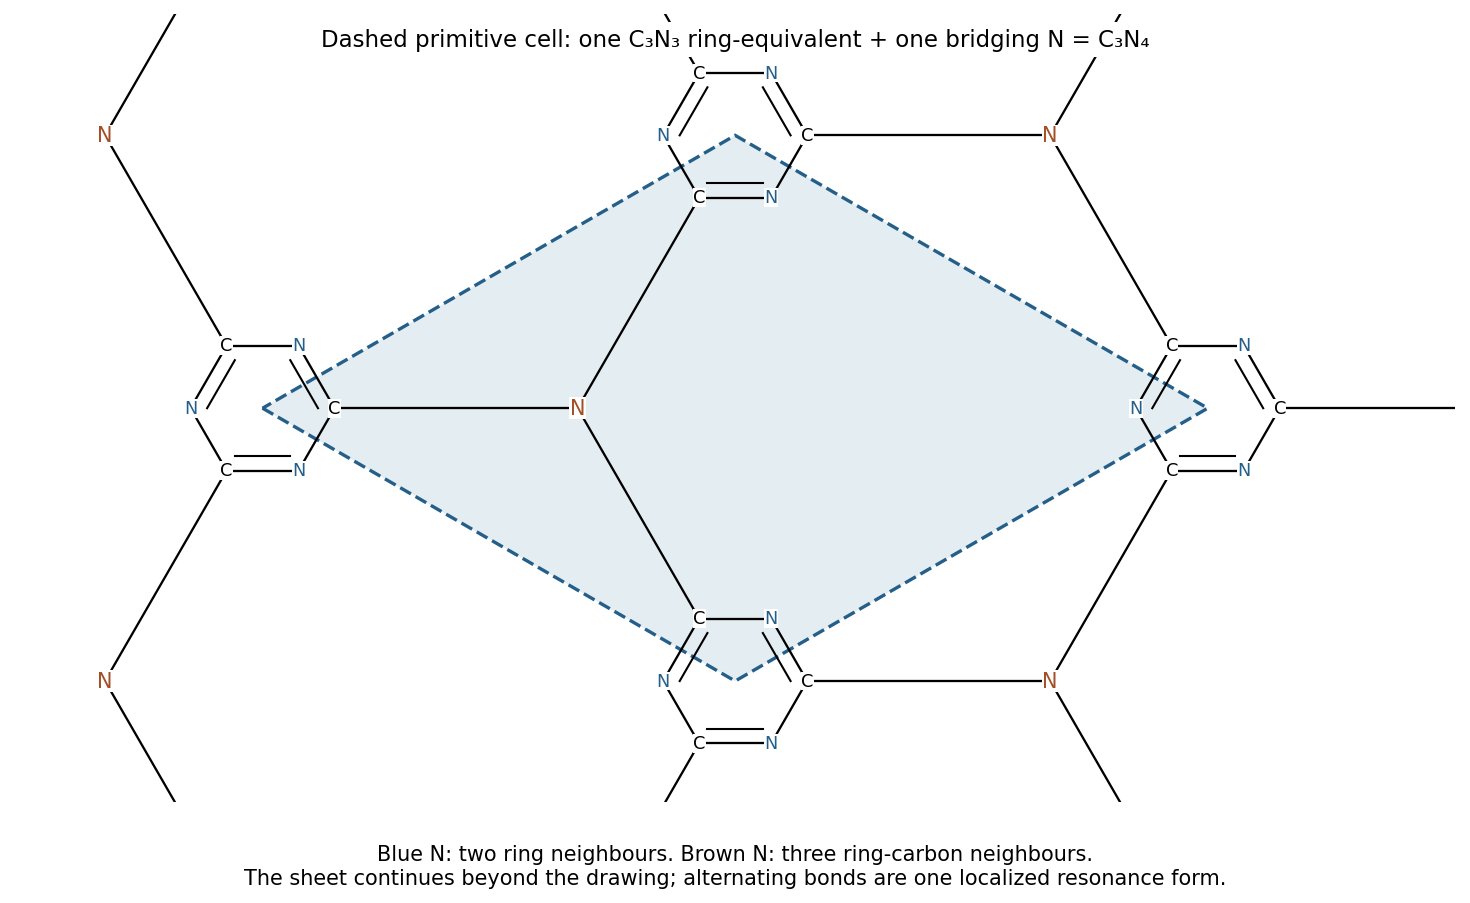

Each triazine ring contributes and has three outward carbon links. A bridging N is shared among three rings, so each ring effectively contributes three thirds of a bridging N: one additional N. The repeat composition is therefore . The highlighted primitive cell contains one ring-equivalent and one bridging nitrogen; the drawn bond alternation is one localized resonance representation.
This is a possible structure, not a unique identification from the empirical formula. Primary synthesis and structural work established triazine-based graphitic carbon nitride as one realizable network. Other commonly prepared carbon nitrides contain heptazine motifs, incomplete condensation and hydrogen, so the name alone should not be treated as an exact structural determination of this decomposition product.[ref]
Three routes are donor doping, acceptor doping and above-gap illumination. All increase mobile carrier populations if the added states or excitations are effective rather than strongly trapped.
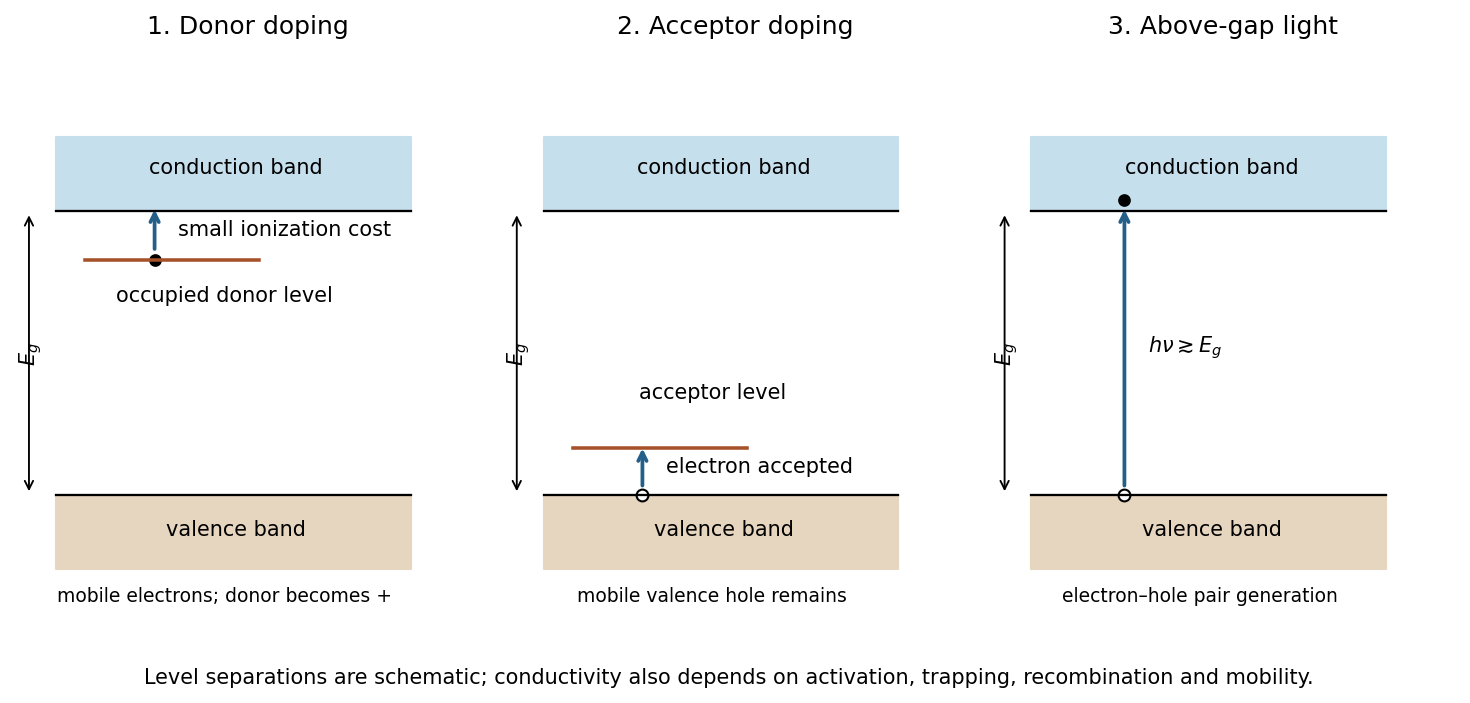
Original band diagrams for donor doping, acceptor doping and photogeneration in a gapped material
. Details. 
An appropriate donor impurity provides an occupied level just below the conduction band. Thermal ionization requires the smaller donor binding energy rather than the full host gap, supplying mobile electrons and n-type semiconductor behaviour. An appropriate acceptor impurity creates an available level just above the valence band; accepting a valence electron leaves a mobile hole, giving p-type semiconductor behaviour. Doping concentrations and compensation must permit useful carriers; an arbitrary impurity need not be an effective dopant.
For photoconductivity, absorbed photons with can excite valence electrons into the conduction band, producing electron–hole pairs. For the stated gap, the nominal threshold is about nm. Optical selection rules and absorption strength can shift the practical onset. The conductivity increase persists only while generation maintains carriers against recombination and trapping; it is not necessarily a permanent change to the material.
In each case, connects increased carrier density to conductivity. Defect-induced reductions in mobility can offset some of the gain, so the diagrams specify mechanisms rather than guarantee a numerical improvement for an unspecified preparation.
Articles by others on the same topic
There are currently no matching articles.