The first transformation is an Achmatowicz rearrangement, followed by protection and selective reduction; the last is a Johnson-Claisen rearrangement.
- Butyllithium removes the furan 2-proton, giving a 2-lithiofuran. Its carbon attacks the aldehyde carbonyl; protonation of the resulting alkoxide gives the furfuryl alcohol. Carbon-carbon bond formation occurs at the lithiated position.
- Electrophilic oxygen transfer from mCPBA to the electron-rich furan gives an oxidized, ring-opened intermediate. Opening the oxygenated furan gives a hydroxy enedicarbonyl: the original furfuryl hydroxyl then attacks the remote aldehyde intramolecularly. This expands the five-membered furan into a six-membered lactol containing an enone carbonyl. The original furfuryl oxygen becomes the pyran ring oxygen; oxidation of the furan supplies the new carbonyl and lactol functionality.
- AgO/MeI converts the lactol OH into its methyl acetal: oxygen attacks methyl iodide by SN2 reaction, with iodide removal and proton transfer. This protects the anomeric position.
- LiAlH delivers hydride to the remaining ketone carbonyl, followed by protonation. The methyl acetal survives these basic reducing conditions, leaving the depicted allylic alcohol.
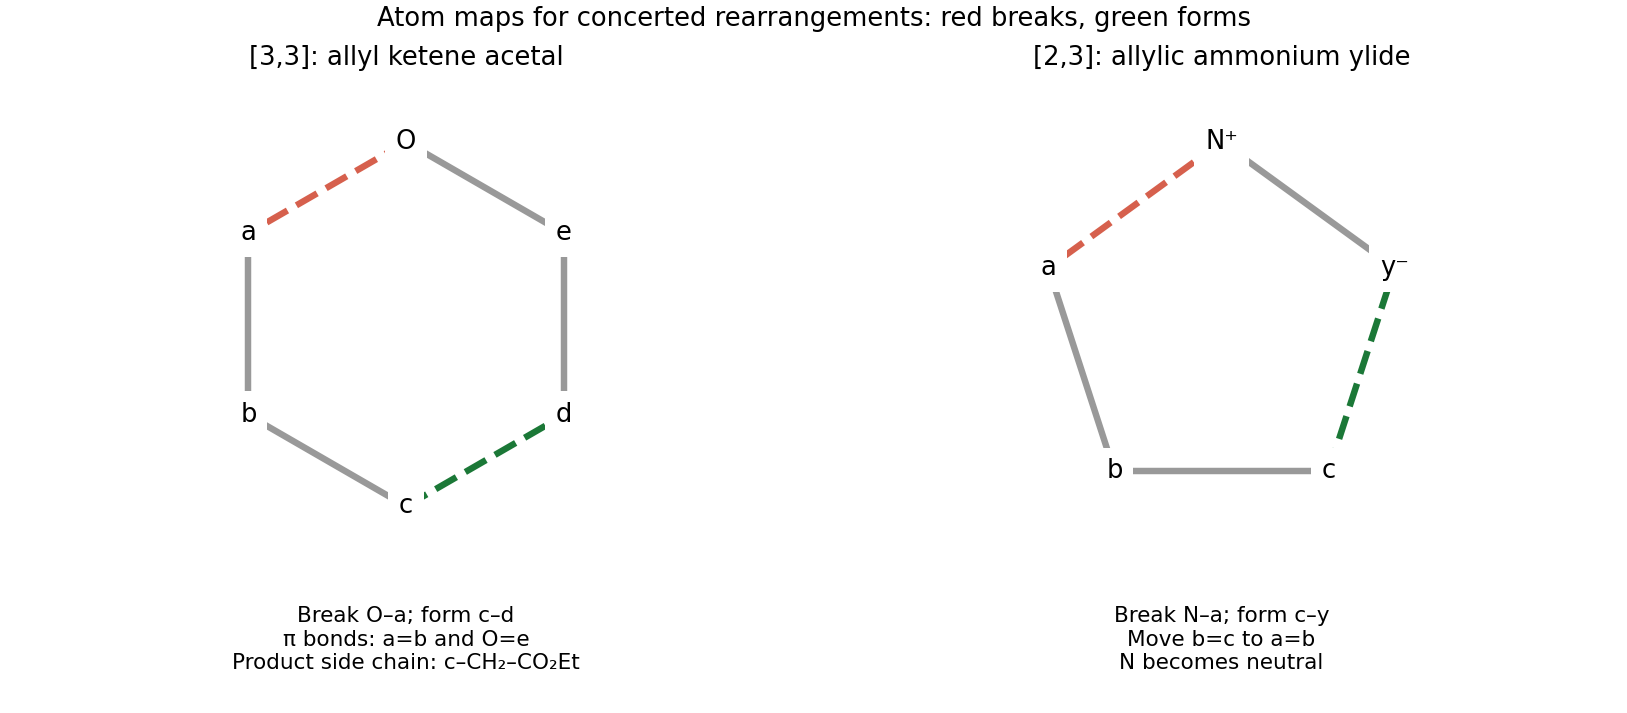
- Under weak-acid catalysis, that allylic OH replaces an ethoxy group of triethyl orthoacetate. Loss of another ethanol molecule, with removal of a methyl proton, gives the mixed ketene acetal . A six-electron, chair-like Claisen rearrangement breaks the allyl-O bond, forms a bond between the terminal allyl carbon and the ketene-acetal CH, and moves the ring double bond one position. Carbonyl formation gives the substituent.
The acetal protects the other oxygenated position during hydride reduction. The orthoester transfers an acetate-derived two-carbon unit through a ketene acetal; it does not directly substitute the allylic OH with an intact acetate ion.

For a concrete choice, draw the intermediate in the planar ring orientation used in the question and put the allylic OH on the upper face, with a solid-wedge C-OH bond. Fix either configuration at the R-bearing and methoxy-bearing centres and retain it throughout; no absolute configuration is specified by the question.
Draw the six-membered rearrangement array with its allylic C-C-C segment along the same ring edge and the O-ketene-acetal tether above that edge. Suprafacial transfer forms the new ring-carbon–CHCOEt bond on that upper face: draw it as a solid wedge, replacing the original allylic OH with the shifted double bond. The R-bearing and methoxy-bearing stereocentres retain their configurations. The opposite OH epimer transfers the tether from the opposite face and gives the opposite new stereocentre. Favor the chair-like transition structure that puts the larger tether substituents outward. This explicitly correlates the selected OH face with the new bond rather than invoking a freely rotating allylic carbocation.
Propionic acid is a weak, relatively high-boiling acid: its normal boiling point is about , compared with about for acetic acid. It provides mild proton catalysis while remaining in the hot orthoester mixture more effectively. This is useful because ketene-acetal formation and the thermal Johnson-Claisen rearrangement require heating, whereas strong acid would threaten the pyran acetal.
This is a practical catalyst choice, not an absolute mechanistic prohibition on acetic acid; acetic-acid-catalysed orthoester rearrangements are also possible.
Articles by others on the same topic
There are currently no matching articles.