Student suggested answers for the past ten years are available on Chemistry Moodle. Official course guide: past papers and suggested answers New solutions for years covered by that archive are skipped; existing wiki drafts are retained.
is nido; and are arachno. The initial hydride removes an acidic bridging hydrogen as a proton, releasing and retaining the potassium counterion. Addition of one unit expands the boron framework; final protonation supplies the neutral hexaborane and KCl. These assignments conserve all the atoms in the displayed sequence, without inserting an unshown dehydrogenation step.
For , take one terminal B–H bond per vertex and remove its electrons from the valence total. The skeletal electron count is . ThusWade's rules assign , and pairs to closo, nido and arachno frameworks respectively. A derives from a six-vertex closo octahedron by removal of one vertex, giving a square-pyramidal boron skeleton. B and C have six-vertex open frameworks formally derived from an eight-vertex closo polyhedron by removing two vertices. Their additional hydrogens are distributed over bridging and other open-face positions; counting every hydrogen as an independent terminal B–H vertex would give the wrong classification.
The arachno assignment of has specific theoretical and NMR support. It should not be replaced by a permanently localized, unrearranged picture: the corresponding symmetric adduct participates in fluxional rearrangement rather than being the lowest-energy structure.[ref]
The Structure of arachno-B6H11-, at -25 degrees C in (CD3)2O, Is Resolved via the ab Initio IGLO-GIAO NMR Procedure by Matthias Hofmann and others. 2000, abstract: arachno minimum, bridging-hydrogen removal from B6H12 and alternative-adduct comparison
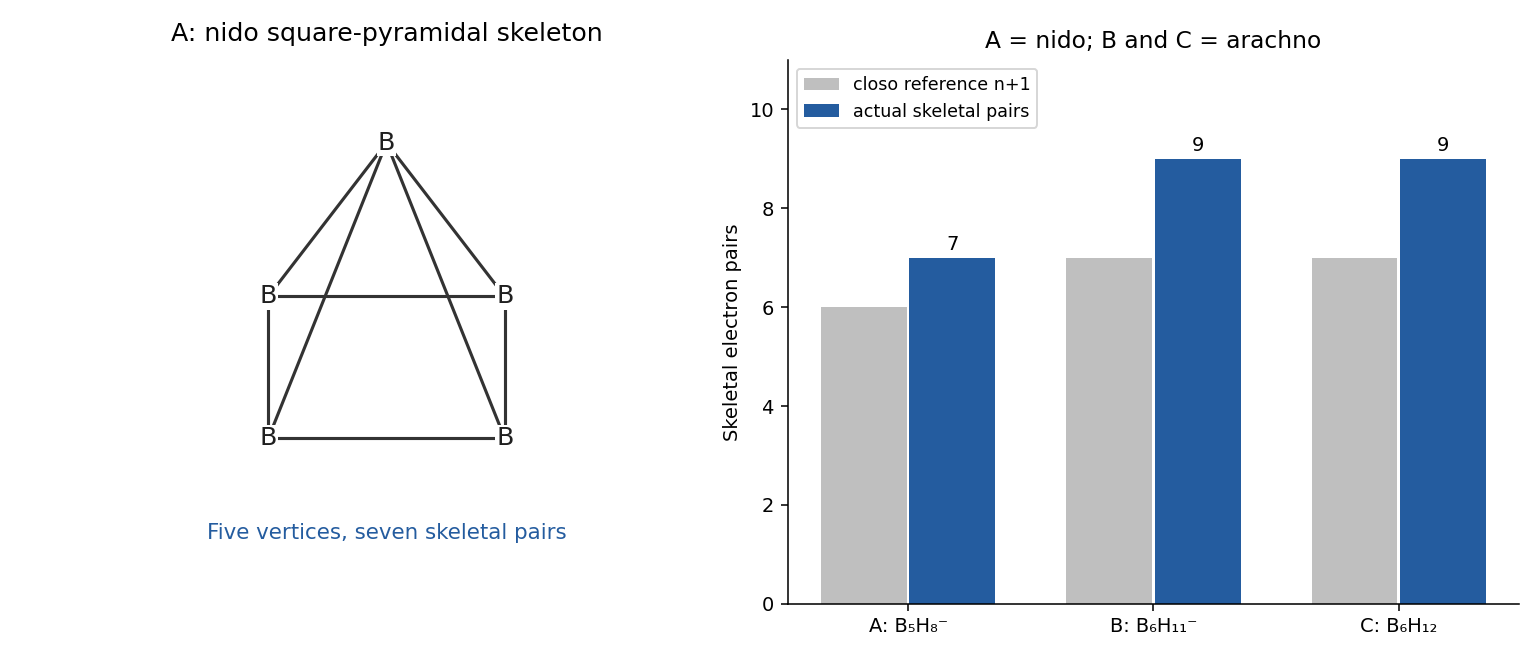
C, the neutral arachno , is the most plausible proton donor among A, B and C. Loss of an open-framework bridging proton produces the experimentally accessible, charge-delocalized conjugate base . In contrast, deprotonating A or B would build a dianion and incurs additional electrostatic cost. The neutral, open hydrogen-rich cluster therefore has a more favourable route to a monoanionic conjugate base. This is a qualitative acidity prediction for the listed species, not a numerical solvent-independent or a claim that all B–H hydrogens are equally acidic.
is approximately square planar; is a closo trigonal bipyramid. Both have 22 valence electrons, and , but their different local orbital requirements prevent inferring identical shapes from this total alone.
For the sulfur ring, four S–S sigma bonds use eight electrons and four in-plane lone pairs use another eight, leaving six electrons in the perpendicular cyclic pi system. The resulting filled bonding and degenerate nonbonding pi levels give a six-pi-electron aromatic description of a four-membered ring, consistent with equivalent S–S links in the ideal square. The out-of-plane p orbitals are retained; this is not a tetrahedral four-vertex closo cage.
For tin, allocate one mainly s-rich lone pair to each atom. The remaining electrons form six skeletal pairs, exactly for . Wade's rules therefore predict a trigonal bipyramid, with three equatorial vertices and two axial vertices. The axial pair is not joined by a polyhedral edge; its bonding is part of the delocalized three-dimensional framework. The contrast is a planar aromatic sigma/pi ring versus a three-dimensional multicentre-bonded Zintl cluster, rather than cationic charge alone imposing planarity or anionic charge alone imposing a polyhedron.
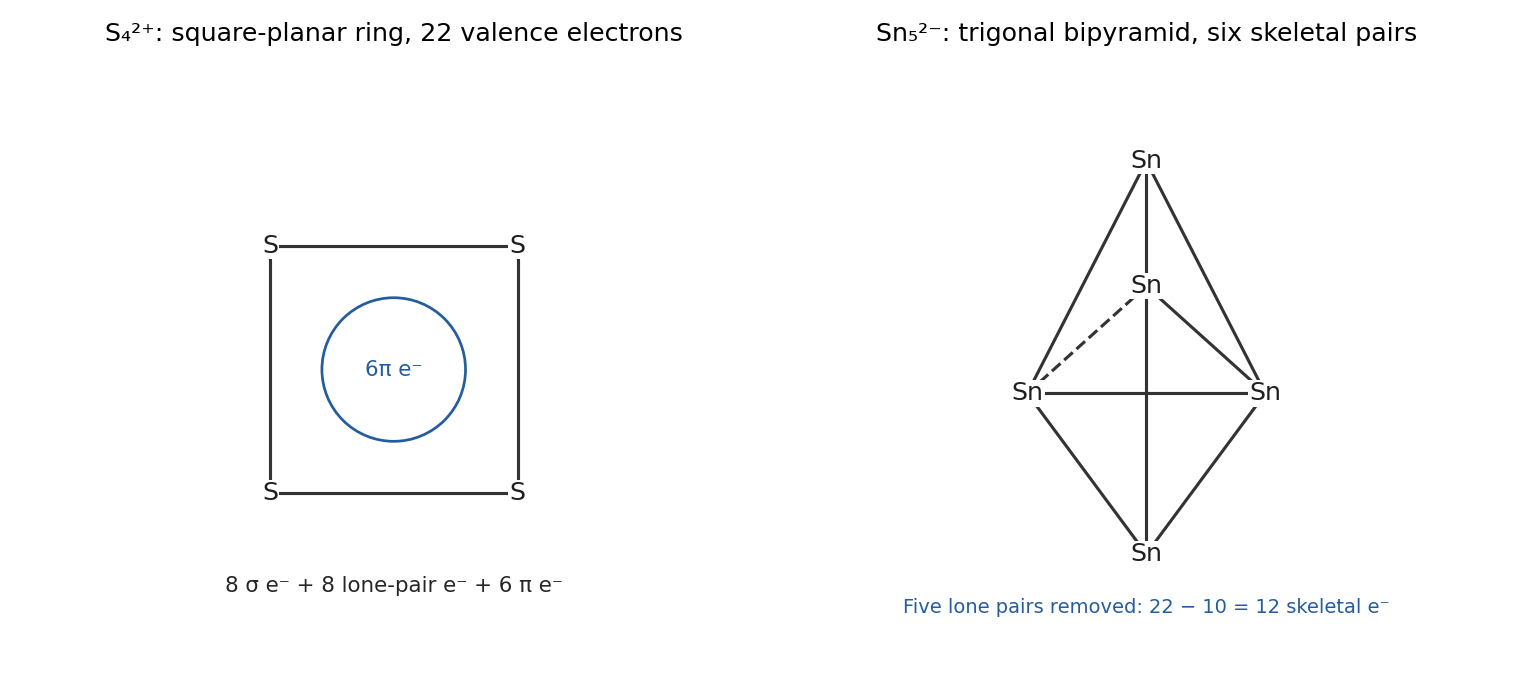
Original square-ring and trigonal-bipyramidal models with their different allocations of the same 22 valence electrons
. Details. Solid contains octahedral clusters, while - has paired molybdenum centres in a layered chloride lattice. The first material is better represented by than by isolated bent or linear molecules. Eight inner chlorides cap the triangular faces of an octahedral metal core, giving . Outer chloride coordination completes the solid: one useful connectivity notation is , with two terminal outer chlorides and four intercluster-shared outer chlorides per octahedron. The empirical total is chlorides for six metals.
Each Mo(II) is formally , giving 24 metal-based electrons. These support extensive Mo–Mo bonding along the 12 octahedral edges; they are not localized as four free spins on each molybdenum. In -, Mo(III) is in edge-sharing octahedral layers. The metal honeycomb distorts into short Mo–Mo pairs. A localized dimer picture uses the six d electrons to fill one sigma and two pi bonding combinations, producing a formal triple bond and pairing the spins. The bulk crystal remains an extended chloride lattice, rather than a collection of unconnected molecules.
By comparison, has predominantly localized high-spin Cr(II), , in Jahn–Teller-distorted chloride octahedra; has layered Cr(III), , with three locally unpaired electrons. Their magnetic exchange is not evidence for the same strong short multiple bonds as in the molybdenum solids. The greater radial extent of Mo orbitals enables larger metal–metal overlap than the more contracted Cr orbitals. Real structure and magnetism still depend on ligand geometry and phase, so the orbital comparison is a trend rather than a prohibition of chromium–chromium bonds in other compounds. Diffraction and magnetic/spectroscopic studies directly distinguish the dimerized - lattice from .[ref]
Structural and Magnetic Transitions Caused by Dimer Formation in the CrCl3–MoCl3 Solid Solution by Samuel Froeschke and others. Introduction and spectroscopic discussion: paired Mo centres, localized Cr spins and diffraction/magnetic evidence
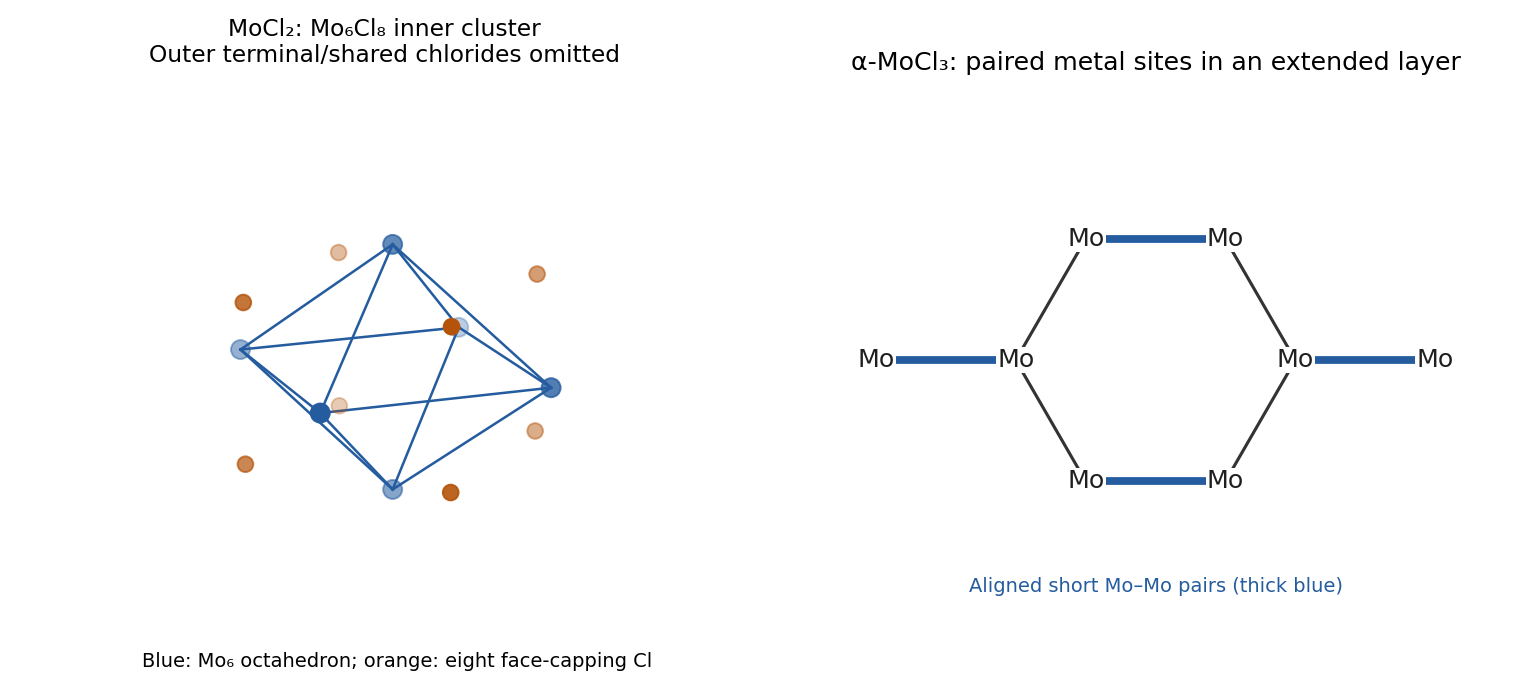
Original Mo6 octahedral-core and paired-layer schematics; selected chloride connectivity is explained in the text
. Details. An alpha decay decreases mass number by four and atomic number by two; a beta-minus decay keeps the mass number fixed and increases atomic number by one. ThereforeThe beta-minus step converts a neutron to a proton with emission of an electron and an electron antineutrino. It is not capture of an orbital electron, which would change the atomic number in the opposite direction.
Radiation changes the solution's chemistry by radiolysis; it does not turn Am(VI) nuclei into Am(V) nuclei. An emitted alpha particle deposits energy into many water and solute molecules, generating ionized/excited species, hydrated electrons, hydrogen atoms and molecular products including hydrogen peroxide. Reducing products can supply the electron needed forFor example, hydrogen peroxide can be oxidized while reducing two americyl(VI) ions:The mM loss of Am(VI) equals the mM appearance of Am(V), preserving the measured dissolved americium total while transferring of electron equivalents. Oxidizing radiolysis products are also present, and the net reduction depends on acid, dissolved gases and scavengers; not every radiolytic event is reducing. In nitric acid, primary kinetic work identifies hydrogen peroxide and radiolytically produced nitrous acid as important reductants, with a rate controlled by total americium activity rather than necessarily by the instantaneous Am(VI) concentration.[ref]
Kinetics of the Autoreduction of Hexavalent Americium in Aqueous Nitric Acid by Travis S. Grimes and others. 2017, abstract: oxidation-state mass balance, UV–visible measurements and radiolytic reducing agents
Let . The useful disconnections are benzoate formation, carbonyl reduction and thiol conjugate addition:The sulfur-bearing carbon is the enone beta carbon, so this is a Michael disconnection, not attack of thiophenol on the alcohol-derived central carbon. Reduce the saturated ketone with sodium borohydride after installing SPh, then benzoylate the alcohol with benzoyl chloride and pyridine or a tertiary amine.
Disconnect the enone to and formaldehyde. Kinetic deprotonation on the methyl side of the hindered unsymmetrical ketone, trapping with formaldehyde, and dehydration gives . This order leaves the substituted side unchanged. An alpha-methylenation/Mannich-base-elimination variant is an equivalent way of installing the terminal enone.
Finally, disconnects by alpha alkylation to butan-2-one and 1-iodopropane. Use the more substituted, ethyl-side enolate of butan-2-one for this step, rather than its usual methyl-side kinetic enolate; an equilibrated/regioisomerically selected enolate or enol derivative is required, and the desired monoalkylated ketone should be isolated. Three carbons from propyl iodide and one from formaldehyde increase the original four-carbon ketone to the required eight-carbon alcohol/thioether skeleton. Choosing ethyl iodide here would leave the target short by one carbon.
This is a constitutional synthesis: the target drawing specifies no configuration at either newly formed stereogenic site, so no unproved stereoselective reduction or alkylation is needed. Regioisomeric enolate alkylation and competing overalkylation are the practical selectivity issues to control, rather than something that follows from writing “base”.
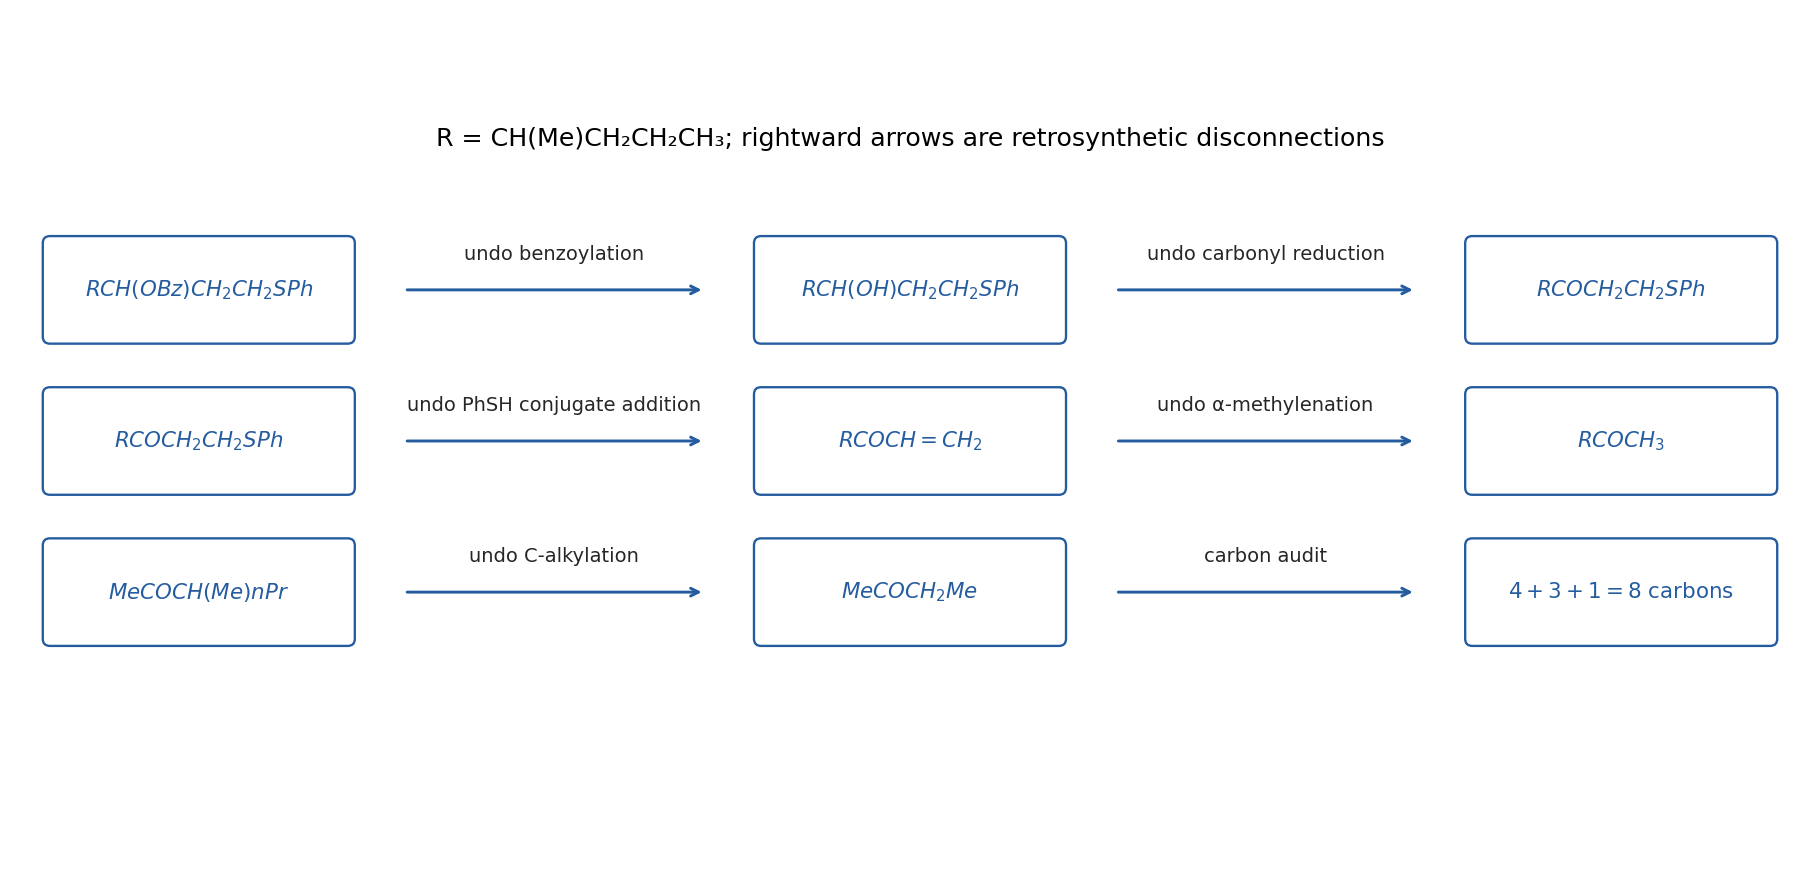
Original atom-counted retrosynthetic sequence from the benzoate/thioether target back to butan-2-one
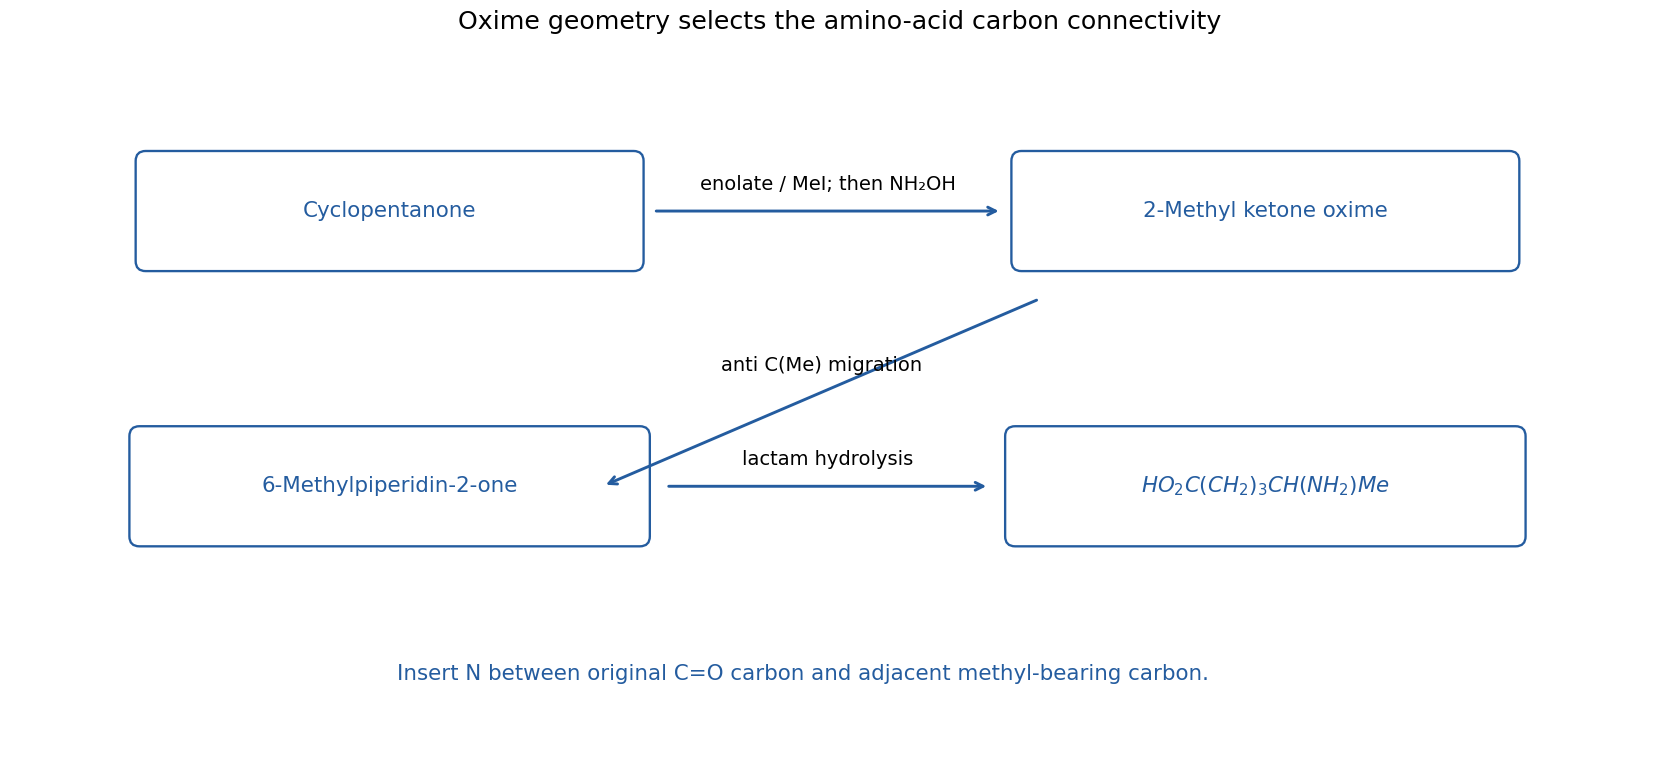
. Details. The target is 5-Aminohexanoic acid, . Disconnect its terminal amine/carboxylic-acid relationship through hydrolysis of 6-methylpiperidin-2-one. This six-membered lactam follows from a regioselected Beckmann ring expansion of 2-methylcyclopentanone oxime.
Start with cyclopentanone. Form its enolate with lithium diisopropylamide in dry THF at low temperature, add methyl iodide, then quench to obtain 2-methylcyclopentanone. Control the methyl iodide/base stoichiometry to suppress a second alkylation. React the ketone with hydroxylamine to form its oxime and select the geometrical oxime isomer in which the bond to the methyl-substituted adjacent ring carbon is anti to the departing oxime oxygen group.
Activate the oxime, for example under appropriate acid-promoted Beckmann conditions. The anti methyl-bearing C–C bond migrates onto nitrogen as the N–O-derived leaving group departs, inserting nitrogen between the original carbonyl carbon and the methyl-substituted ring carbon. Hydration/tautomerization gives 6-methylpiperidin-2-one. The opposite oxime geometry would insert nitrogen on the other side and ultimately place methyl at a different position in the amino acid; oxime geometry is therefore an essential part of this proposal.
Hydrolyse the lactam with aqueous acid and heat, then adjust the work-up to the desired acid/ammonium form. Cleavage of its acyl–nitrogen bond opens the ring to after suitable neutralization. All five original cyclopentanone carbons are retained, and the one methyl carbon comes from methyl iodide. A racemic mixture product is appropriate to the stereochemically unspecified target. The reaction sequence is an organic ring-expansion synthesis and requires no biological transformation.


Open the cyclic ester at its acyl–oxygen bond. The resulting delta-hydroxy acid isOxidation-state reversal at the benzylic alcohol gives . This identifies the key Michael disconnection between the central and groups:The nucleophile is the enolate or enamine of the phenyl ketone. The electrophile's propyl substituent must be alpha to its ester carbonyl; using a beta-propyl acrylate would move the substituent to the wrong ring position.
Disconnect further to the indicated acetophenone. Crossed aldol addition to acetone followed by dehydration gives ; selective conjugate reduction or controlled alkene hydrogenation then gives . This avoids requiring an efficient SN2 alkylation with a secondary isopropyl halide. The alpha-propyl acrylate can be obtained by alpha-hydroxymethylation/dehydration of ethyl pentanoate, , using formaldehyde.
In the forward direction, perform the Michael coupling, reduce the phenyl ketone selectively with sodium borohydride while retaining the ordinary ester, hydrolyse the ester, and cyclize the delta-hydroxy acid by controlled acid-catalysed lactonization or an activated-acid coupling. The six-membered ring then has the order carbonyl–––––O. This atom order checks all three substituent positions. No relative configuration is specified, so the scheme does not imply a single diastereomer from the Michael addition or reduction.
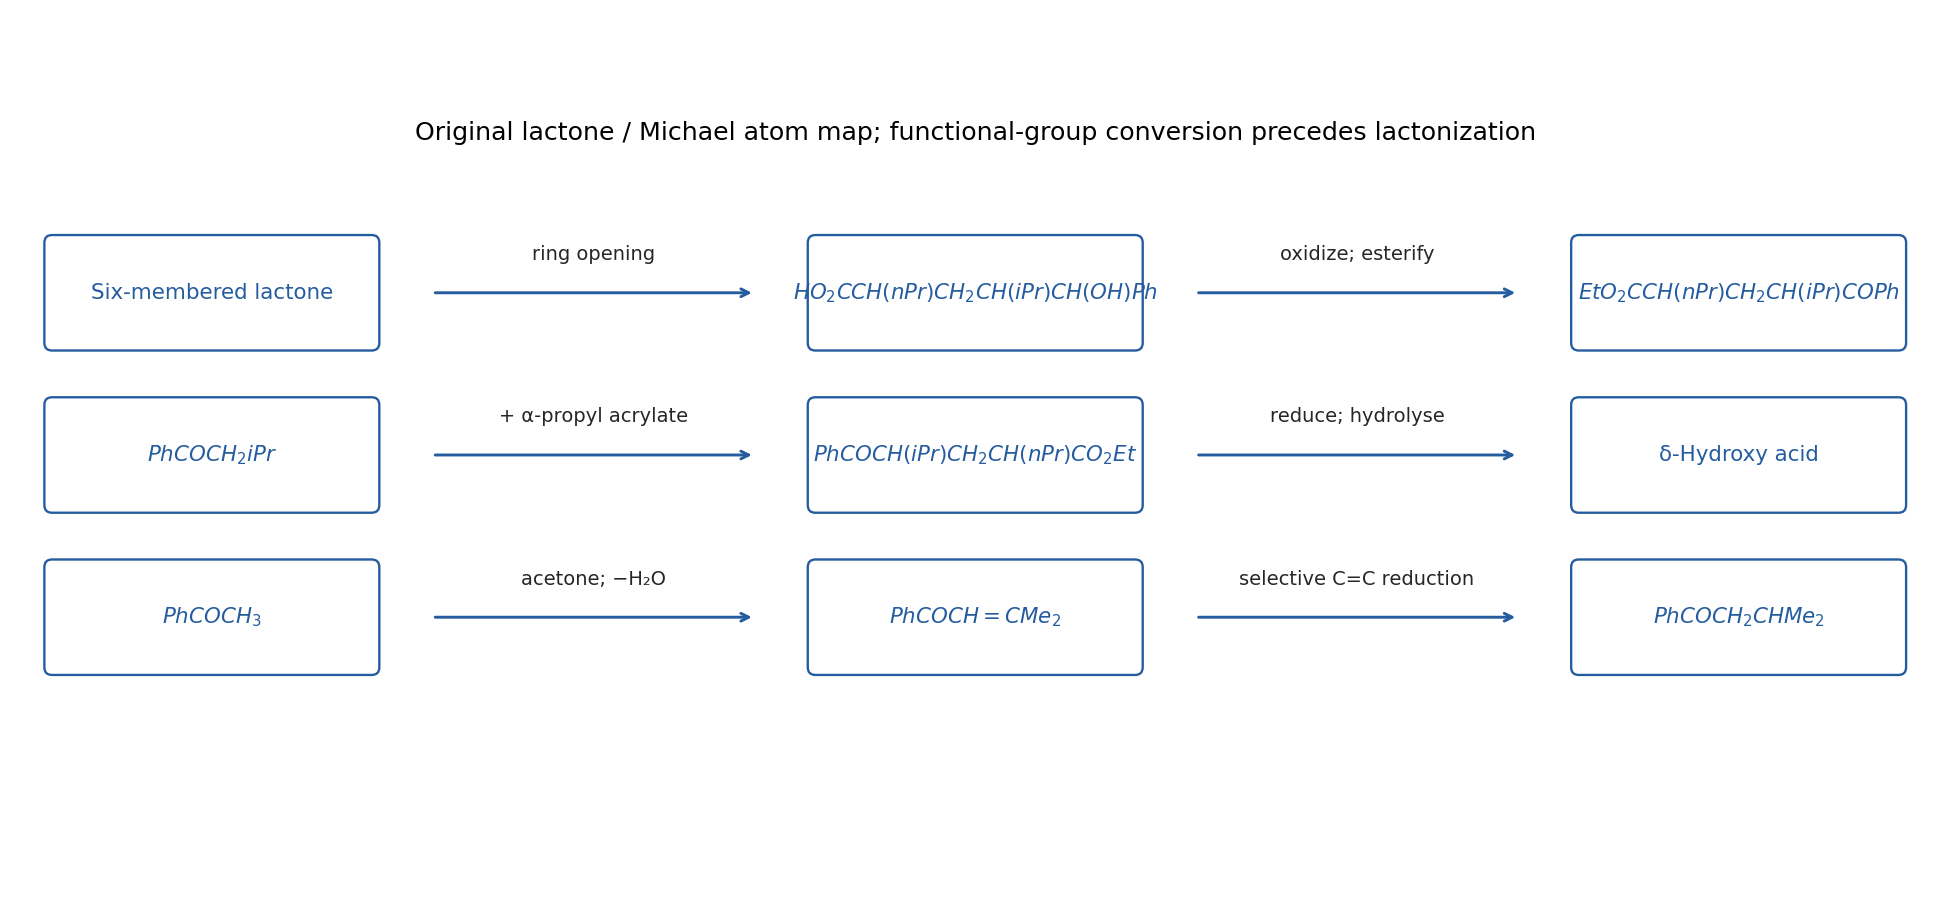
Original lactone-opening and Michael disconnections, with propyl and isopropyl positions tracked separately
. Details. 
Disconnect the tertiary amide into cyclohex-3-ene-1-carboxylic acid and the secondary amine . The acid-derived ring is an immediate Diels–Alder disconnection: buta-1,3-diene plus the indicated methyl acrylate gives methyl cyclohex-3-ene-1-carboxylate. Heating, or suitable Lewis-acid activation of the dienophile, forms the two new sigma bonds and leaves the diene's internal double bond as the ring alkene.
Hydrolyse this ester to the acid. Convert it to its acid chloride with oxalyl chloride or thionyl chloride, and react with the secondary amine in the presence of a base to capture HCl. The tertiary amide now contains the complete cyclohexene acyl fragment; direct conjugate addition of the amine to methyl acrylate before ring construction would give the wrong connectivity.
Prepare the amine by reductive amination of 4-phenylbutan-2-one, , with ethylamine, using sodium triacetoxyborohydride or another iminium-selective reducing system. This places nitrogen on , not on a terminal methylene. If a route from simpler carbon compounds is wanted, crossed aldol condensation of acetone with benzaldehyde gives , and selective C=C reduction gives the required ketone before reductive amination.
The acid branch retains the acrylic carbonyl as the amide carbonyl; the amine branch supplies both N-ethyl and N-(4-phenylbutan-2-yl) substituents. The unlabelled stereogenic centres may be racemic mixture. This sequence separates ring formation from amide formation and avoids invoking uncontrolled alkylation of a tertiary amide nitrogen.
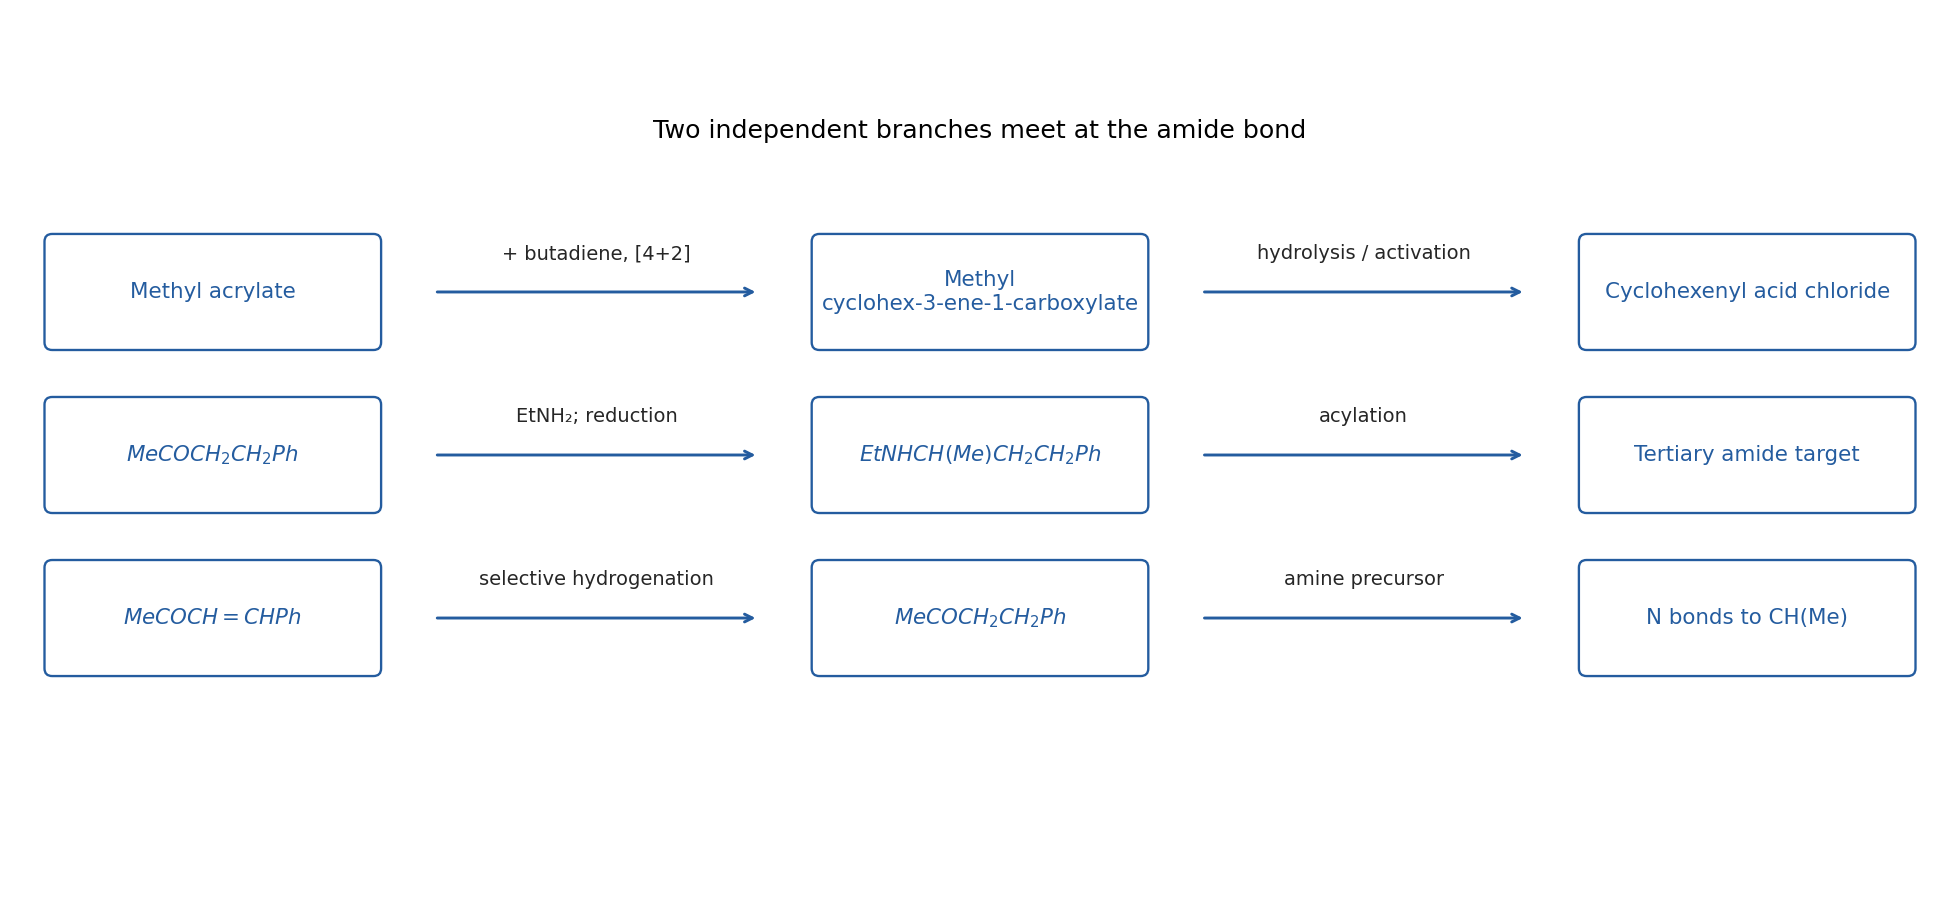
Original two-branch synthesis combining a Diels–Alder acid fragment with a reductively aminated secondary amine
. Details. 
An isolobal analogy compares fragment frontier orbitals, not atomic sizes or total electron counts. The neutral BH fragment has four valence electrons; two make the external B–H bond, leaving two skeletal electrons in three frontier orbitals directed towards possible cluster neighbours. For the neutral fragment, contributes five electrons and cobalt nine, giving 14. Removing the 12 electrons assigned to filled, predominantly non-skeletal fragment orbitals again leaves two electrons in three cluster-directed frontier orbitals.
The corresponding frontier sets have comparable numbers, spatial symmetries and occupations, allowing replacement of a BH vertex by CpCo in a suitable cluster-electron count. This is why the metal fragment can cap an open carborane face without adding an arbitrary new set of skeletal electrons. Exact orbital energies, ligand preferences and bond lengths need not be identical.[ref]
Building Bridges Between Inorganic and Organic Chemistry (Nobel Lecture) by Roald Hoffmann. 1982, isolobal fragment correspondence and its use across organic/inorganic bonding
React pentaborane with an alkyne under relatively mild conditions to obtain a nido framework, conventionally nido-2,3-dicarbahexaborane(8) for acetylene. The two entering carbon atoms initially retain their adjacency. A specific mild example uses 2-butyne with triethylamine-assisted at or below room temperature to give nido-. This is a practical route to an explicitly nido carborane, rather than an assumption that an alkyne supplies a closo cage under every condition.[ref]
Improved Synthetic Routes to Small Carboranes: Bench-Scale Preparation of nido-2,3-C2B4H8 Derivatives from B5H9 and from B3H8- Salts by Narayan S. Hosmane and Russell N. Grimes. 1979, abstract and preparation discussion: amine-assisted substituted-alkyne reaction under mild conditions
More energetic gas-phase reaction or pyrolysis promotes hydrogen loss, boron-fragment loss and cage rearrangement, giving closo carboranes, for example - and -, with other cage sizes possible in the mixture. Thermolysis of a previously formed nido carborane is an alternative route to this stage. The point is not a clean same-composition nido-to-closo isomerization: the change of hydrogen content and sometimes cage size must be retained in the formulas.
Milder conditions favour kinetically trapped adjacent-carbon products; higher temperatures permit migration to carbon-separated arrangements, often increasing favourable B–C contacts relative to adjacent C–C/B–B contacts. Conditions and Lewis bases also influence which separated-carbon cage is isolated; temperature alone does not prescribe a single quantitative isomer distribution. Primary thermolysis studies and the small-carborane literature document both cage rearrangement and reagent-dependent product mixtures.[ref]
Synthesis of Carboranes from Dihydrocarboranes by Thomas P. Onak and others. 1963, printed p. 3378, introduction and product summary
For , twelve BH vertices supply skeletal electrons and the charge adds two:For the cobaltacarborane, nine BH vertices supply 18, two CH vertices supply six and CpCo supplies two by (a):Both are 12-vertex closo icosahedra with 13 skeletal electron pairs. In the cobalt compound, the Cp ring is external to the cage. cobalt caps the five-membered open face of the 11-vertex dicarbollide fragment and is a vertex of the resulting icosahedron, giving the usual adjacent-carbon cobaltacarborane framework. Counting all five Cp carbons as additional polyhedral vertices would be incorrect.
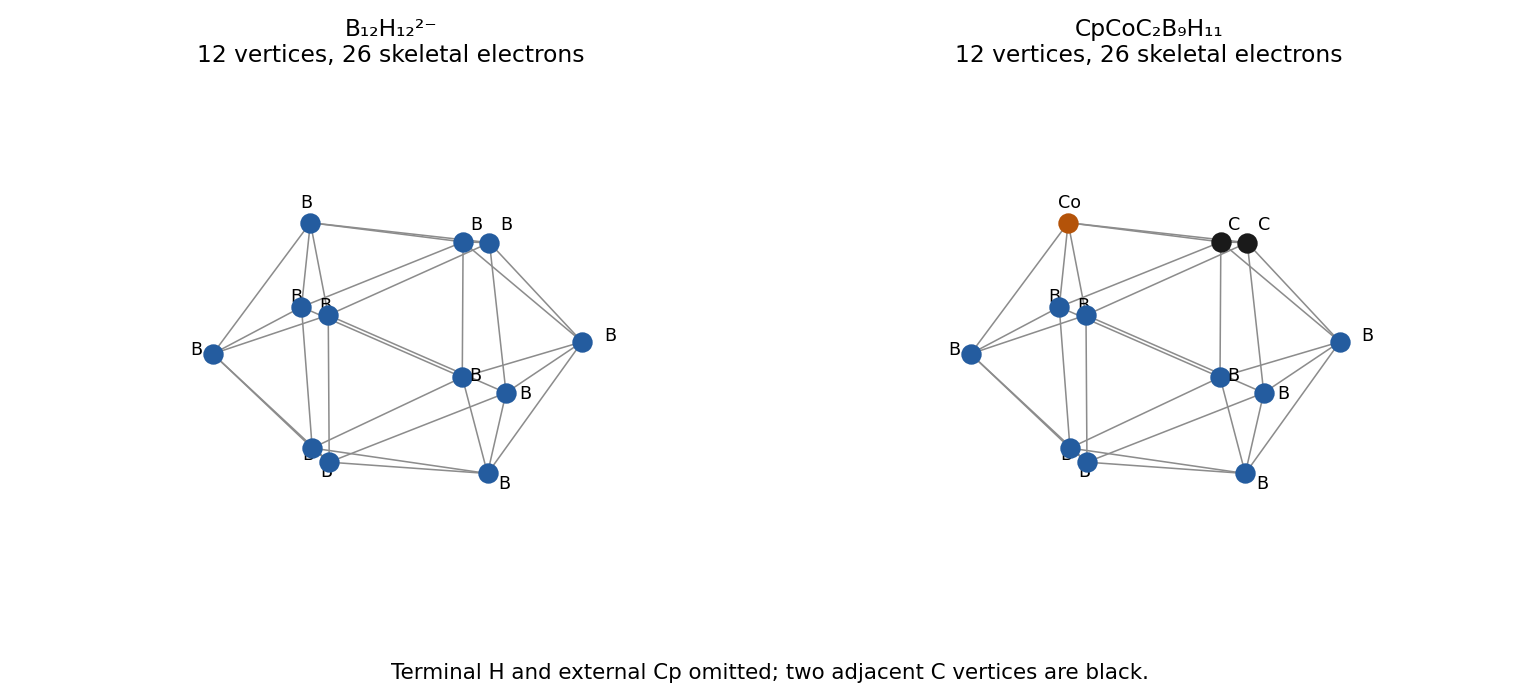
Original 12-vertex closo frameworks for dodecaborate and the adjacent-carbon cobaltacarborane, with external ligands omitted
. Details. 
Write for the open-face dicarbollide ligand. The structure consistent with the stated attachment is , with a direct Mo–Mo connection, three carbonyls on the cage-capping Mo and five on the outside Mo. The first Mo has an interaction with the five-vertex face and completes a 12-vertex metallacarborane cage; the second is exo to that cage. The two tetramethylammonium ions are counterions.
The unsaturated fragment is a 16-electron Lewis acceptor; is an 18-electron metal-base fragment in ionic counting, . Donation from a filled Mo-based orbital supplies the external fragment with a pair. This donor–acceptor representation avoids the mistaken claim that both initially saturated 18-electron fragments simply add a new conventional bond without changing anything.
An alternative formal oxidation-state allocation for a single covalent Mo–Mo bond is cage Mo(I) and external Mo(−I): the local counts are and . These are electron-bookkeeping alternatives to the metal-base/metal-acid picture, not a measurement of integral site charges. With terminal carbonyls, the external group can be approximately octahedral when the Mo–Mo direction occupies its sixth coordination direction. The cage-containing unit must accommodate an exo metal interaction in addition to its facial cage/CO environment; its geometry need not be a perfectly undistorted isolated half-sandwich.
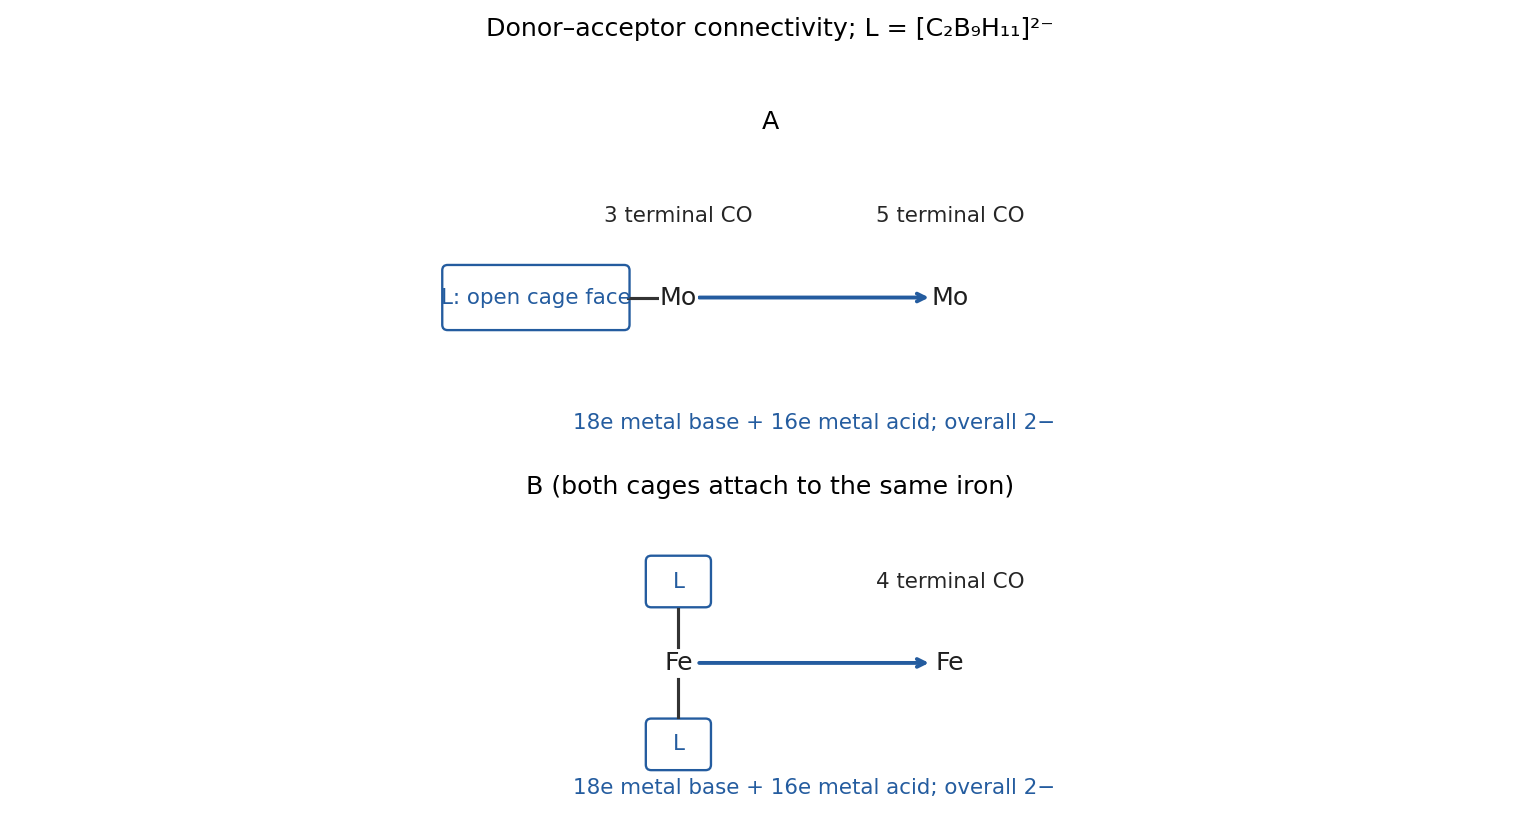
Original donor–acceptor models for the exo Mo and Fe carbonyl fragments attached to cage-bound metal donors
. Details. 
The wording places both dicarbollide cages on the same iron, leaving the second iron outside the cages. The corresponding proposal is , with the first iron sandwiched between the two open faces and the second carrying four carbonyls. It is not the symmetric arrangement with one cage on each iron, which would contradict the specified single cage-interacting metal site.
The donor fragment has electrons; the external neutral fragment has . Metal-to-metal donation therefore provides a natural 18-electron acceptor adduct. In covalent-single-bond oxidation-state bookkeeping, the analogous assignments Fe(III)/Fe(−I) give and . In either account the whole species has two negative charges. The outside iron can have an approximately trigonal-bipyramidal five-coordinate arrangement when the Fe–Fe direction and its four CO ligands are counted.
The important structural distinction is a bis(dicarbollide) iron donor joined to an exo tetracarbonyl iron acceptor. Formula and electron counting alone do not uniquely fix all carbonyl angles or prove a particular crystalline conformation.
Neither formula requires a bridging carbonyl to repair an electron deficiency: the terminal-CO donor–acceptor structures in (d) and (e) already give an electron-counting rationale for the metal–metal interaction. A predominantly terminal-carbonyl model is a reasonable starting proposal for A; bridging or semibridging CO is more geometrically plausible for the shorter Fe–Fe contact in B. Larger metal separation makes a carbonyl simultaneously engaging both Mo atoms less favourable, while carbonyl redistribution or tilting can be easier around a compact Fe pair.
This is a qualified structural prediction, not a determination from molecular formula. A bridging CO can change the relative hapticity and local electron allocation, and the two metal centres are inequivalent; a large frequency shift by itself need not prove a symmetric bridge. Infrared carbonyl frequencies combined with diffraction would test terminal versus bridging or semibridging assignments. The given low-temperature preparation can trap a coordination pattern before complete carbonyl redistribution.
Ultrashort metal–metal contacts are favoured when the valence configuration fills strongly metal–metal-bonding orbitals while leaving their antibonding partners empty. Early or middle transition metals in low oxidation states can retain enough d electrons for multiple bonding without the unavoidable antibonding occupation of a more filled d shell. Good radial and angular overlap, a compatible metal–metal axis, and low competing ligand-field stabilization all matter; a short contact should be interpreted together with an orbital/electron-counting model, not automatically called a high-order bond.
Ligands should support a geometry in which the relevant d orbitals can overlap and protect the low-valent pair from oligomerization or oxidative attack. Bulky aryl-substituted bridging amidinates can provide this protection while maintaining an appropriate small coordination number. Strongly coordinating axial ligands or inappropriate pi interactions can compete with metal–metal bonding, especially its weaker delta components. For quadruple or quintuple bonds, retaining two appropriately oriented delta-capable orbitals is particularly important; a geometry optimized only for metal–ligand bonding can remove that advantage.
Ligand bridges need not literally squeeze two otherwise unbound metals together. Comparing bond lengths across oxidation states and across related ligands helps distinguish electronic multiple bonding from enforced steric compression. The lantern-type dichromium experiment in (ii) is informative because reduction changes the electron occupation within essentially the same three-bridge supporting framework.[ref]
Remarkably Short Metal-Metal Bonds: A Lantern-Type Quintuply Bonded Dichromium(I) Complex by Yi-Chou Tsai and others. 2008, printed pp. 7250–7253: mixed-valent precursor, reduced anion, orbital analysis and discussion of ligand constraints
chromium is the characteristic assignment, giving Cr(I)–Cr(I) with a formal quintuple bond. Each formamidinate is monoanionic, so gives . chromium(I) is and the pair supplies ten d electrons. Relative to a metal–metal axis, the idealized bonding combinations are sigma, pi and delta:End-on sigma overlap gives the largest splitting, pi overlap a smaller splitting, and delta overlap the weakest. A useful qualitative ordering places bonding sigma lowest, then the two pi combinations, then the two delta combinations; all are occupied. Their antibonding partners are empty, with the weakly split delta-star levels nearest their bonding counterparts. The actual three-bridge ligand field can mix and split symmetry-compatible levels, so the diagram is a bond-axis model rather than five perfectly degenerate metal d levels in the molecular complex.
The identified lantern-type anion has an experimentally determined Cr–Cr distance near Å; reduction of its nine-bonding-electron precursor supplies the tenth electron. This corroborates the model more specifically than citing “short bonds” in an unrelated framework.[ref]
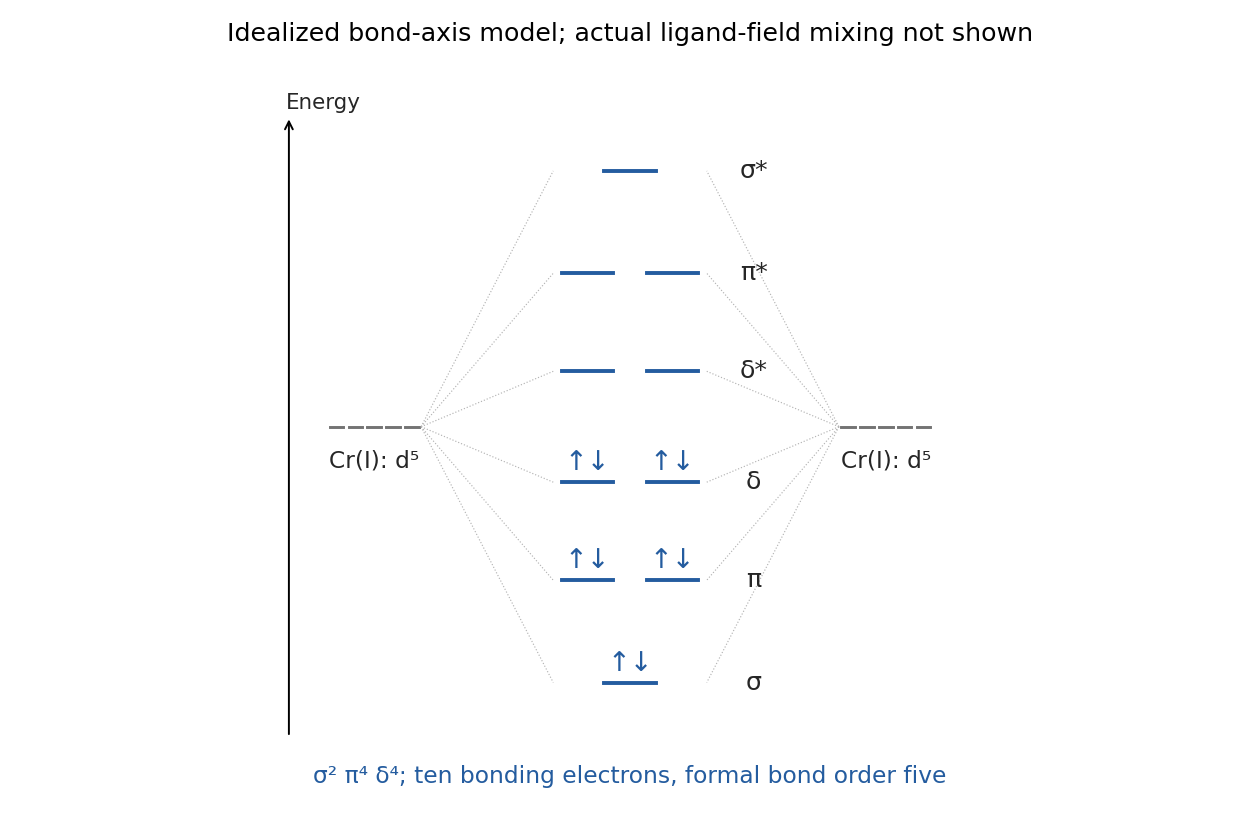
Original idealized d-orbital combinations and ten-electron occupancy for a sigma-two-pi-two-delta quintuple bond
. Details. 
Use the balanced aqueous-acid reactionA Hess cycle first atomizes the metal, then ionizes its gas-phase atoms, then hydrates the triply charged gas ions:Close the reference leg by combining the three electrons with to make . Calling that element-independent contribution givesA single-ion standard enthalpy of formation uses the conventional hydrogen-ion reference; it must not be treated as a balanced creation of net charge from neutral metal without an electron/reference term. The constant hydrogen leg does not affect comparisons along the lanthanide series.
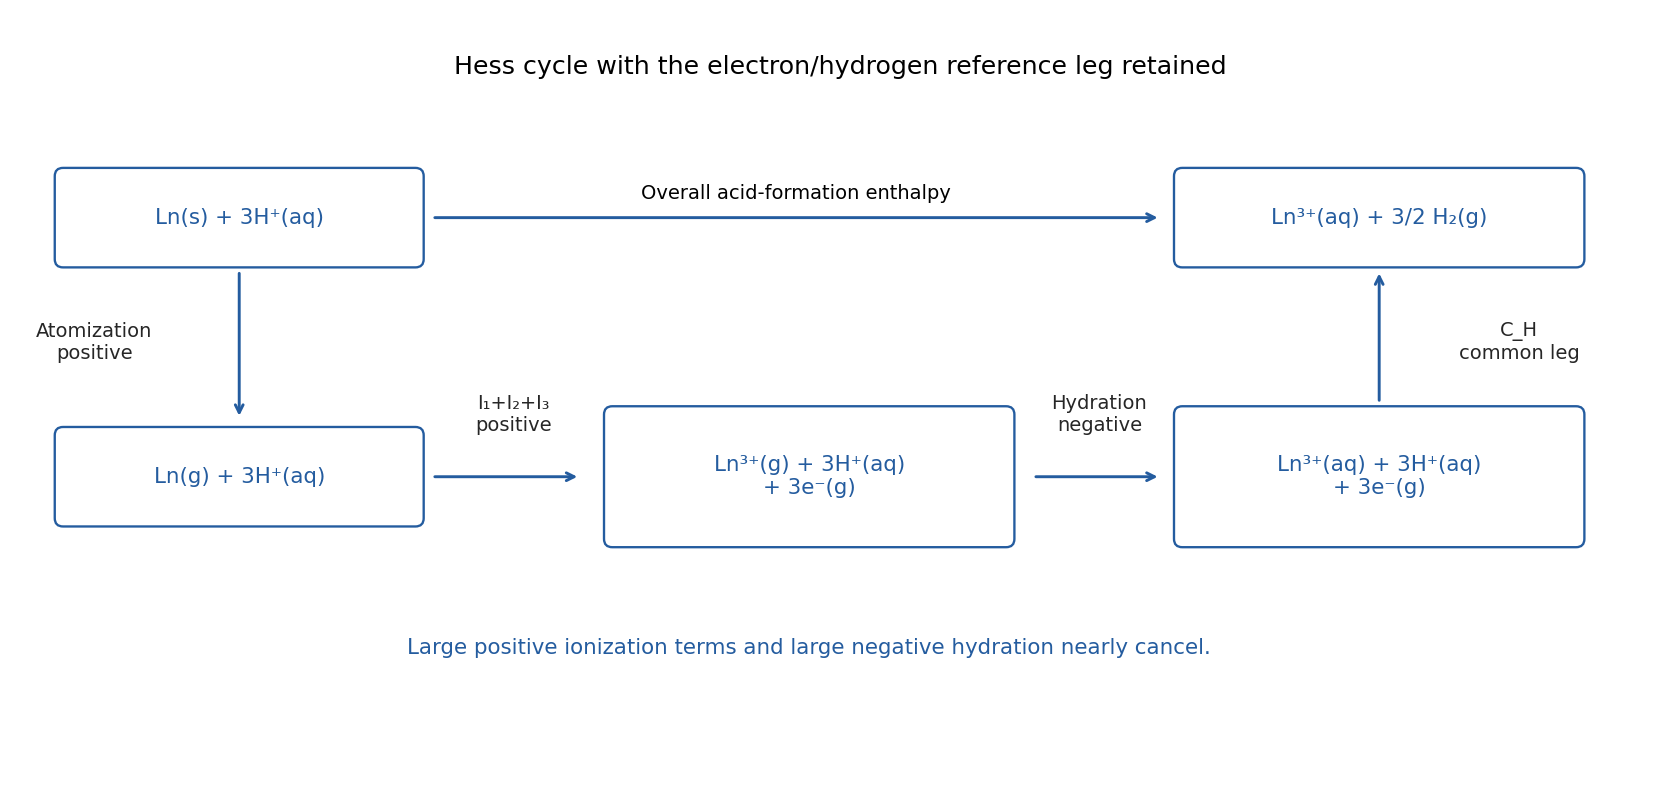
Original Hess cycle separating atomization, three ionizations, hydration and the common aqueous hydrogen reference leg
. Details. 
The large positive sum of ionization energies and large negative hydration enthalpy dominate, with a smaller but significant positive atomization term. Ionization depends on effective nuclear attraction, orbital energies and the stability of successive gas-ion electron configurations; removing the third electron may require disruption of a particularly stable divalent f configuration. Atomization depends on the metal's cohesive bonding and valence participation, and need not vary perfectly smoothly.
Hydration becomes more favourable for higher charge density. All the compared ions have charge , so smaller effective ionic radius generally gives more negative hydration enthalpy. Incomplete shielding by the 4f shell produces the lanthanide contraction and strengthens ion–water attraction along the series. A Born-type electrostatic estimate has a stabilization scale proportional to , but hydration-shell structure, water polarization and temperature dependence prevent treating that crude scale as an exact enthalpy formula. The hydration trend is smoother than the sharp configuration-dependent ionization anomalies. The measured formation enthalpy reflects cancellation of large terms, so comparison of just one term can give a misleading prediction.
The unusually stable divalent configurations Eu(II) and Yb(II) make their third ionization particularly costly. Forming Eu(III), , or Yb(III), removes an electron from a half-filled or filled f shell. The positive ionization-energy contribution therefore increases relative to the smoother background across the series.
Their trivalent ionic radii and hydration enthalpies do not supply a matching abrupt extra stabilization. Differences in atomization and the divalent character of elemental Eu and Yb partly compensate the ionization anomaly but do not erase the net less favourable trivalent aqueous formation enthalpy. This explanation concerns formation of the aqueous ions, not a claim that or is intrinsically the most stable isolated f configuration.
There are three largely independent constructions: a substituted Robinson-annulation core, a protected vicinal diol, and the ester acyl fragment. Disconnect the ester first to an allylic alcohol on the bicyclic ring and the acid . Disconnect the benzylidene acetal to a side chain and benzaldehyde. Reversing 1,2-reduction of the allylic alcohol gives the corresponding bicyclic enone; its six-membered enone ring is a Robinson disconnection back to a suitably alpha-substituted cyclohexanone and methyl vinyl ketone.
An efficient way to protect the side chain before annulation is to prepare 4-(bromomethyl)-2-phenyl-1,3-dioxolane from and benzaldehyde with mild acid and water removal. The desired five-membered acetal protects the adjacent hydroxyl groups, while its external primary bromide remains an alkylating site. Alpha alkylation of the indicated cyclohexanone with this bromide gives a ring bearing in the benzylidene-protected form. There is no need to expose the eventual ring enone to a nonselective late dihydroxylation.
Generate the enolate at the other, less substituted alpha position of the cyclohexanone and add methyl vinyl ketone. Hydrolysis of an enamine-mediated Michael variant or direct enolate trapping gives the appropriate 1,5-dicarbonyl. Intramolecular aldol condensation closes a six-membered ring onto the original cyclohexanone carbonyl. Dehydration gives the fused bicyclic enone, with the protected three-carbon side chain on the adjacent carbon of the original ring. Regioselecting the initial Michael addition is essential: reaction at the side-chain-bearing alpha site would place that substituent at the wrong fusion position.
Reduce the new enone carbonyl by 1,2-addition, for example sodium borohydride with cerium(III) chloride under Luche-type conditions, to retain the displayed ring C=C bond and produce the allylic alcohol. The benzylidene acetal should be maintained through these neutral/basic operations. The target has no wedge/dash assignments, so a constitutional solution does not require a claim of a unique fused-ring/alcohol diastereomer; isolate the required constitutional annulation product.
For the acyl branch, form the enolate of the indicated methyl butanoate, add acetone, then dehydrate the beta-hydroxy ester:Hydrogenate this isolated alkene before coupling it to the bicyclic substrate, giving . saponification gives ; activate it as its acid chloride or use a mild ester-coupling reagent, then acylate the bicyclic allylic alcohol. Performing the hydrogenation on the separate acyl branch avoids reducing the target ring alkene. Strong acid should be avoided during the final esterification because the benzylidene group is acid-labile.
The carbon audit is particularly useful: the fused rings contain six carbons from cyclohexanone plus four from methyl vinyl ketone; the side chain has three carbons from the protected bromodiol; the benzylidene unit supplies its acetal carbon and phenyl group. In the acyl fragment, the four butanoate acyl carbons plus three acetone carbons give seven; the starting ester methoxy carbon is lost during hydrolysis.
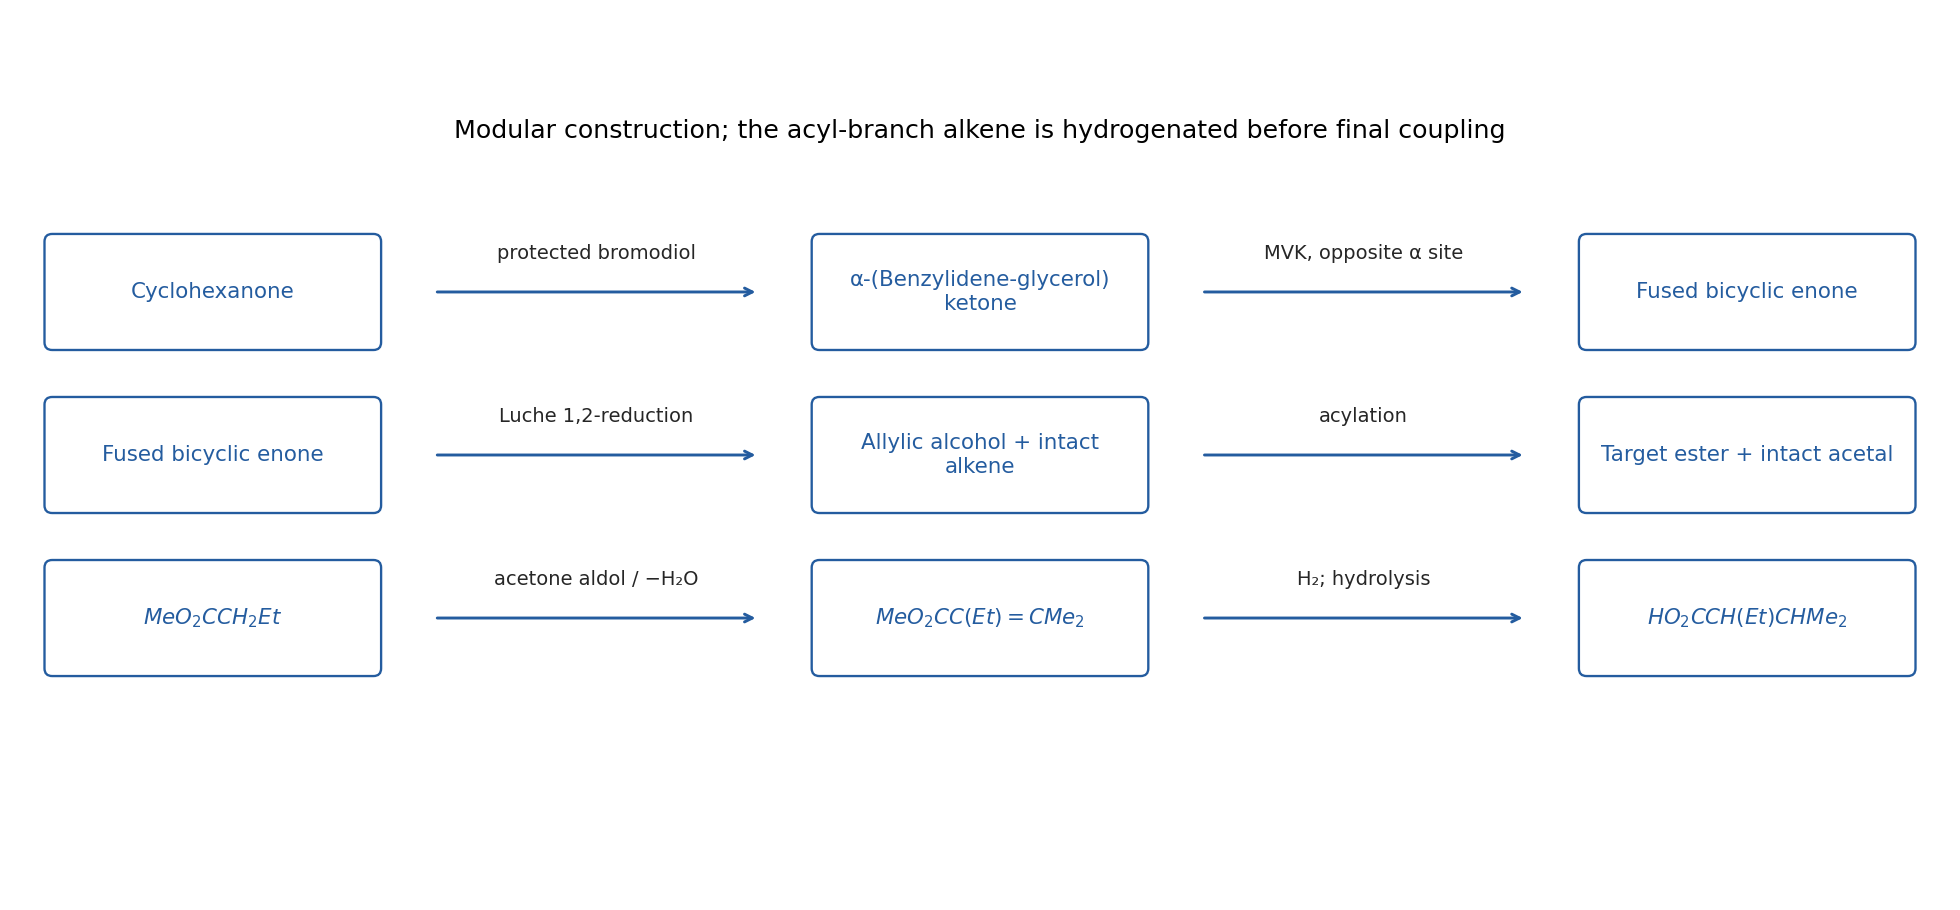
Original modular disconnections separating protected side-chain installation, regioselected Robinson annulation and synthesis of the seven-carbon acyl group
. Details. 
Both side chains have the order ring–, and both printed double bonds have the Z geometry. The two carbon side chains are cis to one another; equivalently the labelled ring H and ring Me are cis on the opposite face. A useful common late precursor therefore has two cis ring– substituents, which can undergo two Z-selective ethylidene Wittig reactions.
Use the starting 2-methylcyclohex-2-en-1-one in a reductive alpha-alkylation. Conjugate reduction under dissolving-metal/enolate-trapping conditions, followed by a primary electrophile , installs a protected two-carbon alcohol side chain at the original methyl-bearing alpha carbon. This gives 2-methyl-2-(2-siloxyethyl)cyclohexanone. The carbon at that position is now quaternary, with its original methyl retained. Alternatively, reduce the enone to 2-methylcyclohexanone and deliberately form/select its more substituted enolate before alkylation; its opposite kinetic enolate would give the wrong regioisomer.
Convert the remaining ring ketone to an exocyclic enoate by Horner–Wadsworth–Emmons olefination with a phosphonoacetate:Hydrogenate this sole alkene to give ring– at that carbon. Select the diastereomer with this substituent cis to the pre-existing protected hydroxyethyl side chain, so the newly installed ring H is cis to ring Me. Less hindered-face hydrogenation can assist this relationship, but the route should include separation/verification of the desired cis product rather than assert complete stereoselectivity without data.
Reduce the ester with lithium aluminium hydride to ring– and remove the TBS group from the other side chain. Both appendages are now two-carbon primary alcohols. Oxidize them to aldehydes under non-overoxidizing conditions, for example Dess–Martin oxidation, obtaining the cis dialdehyde with two ring– groups. Treat with excess salt-controlled, nonstabilized ethylidenetriphenylphosphorane under conditions favouring kinetic Z olefination; isolate the Z,Z product. Each aldehyde carbon becomes the inner vinylic carbon of its crotyl group, while the ylide supplies .
This order postpones both stereodefined alkenes until after hydrogenation, so they are not accidentally saturated. It also avoids assuming that a crotyl grignard reagent adds without allylic transposition or that direct alcohol deoxygenation fixes the required ring stereochemistry. An achiral synthesis gives a racemic mixture pair with the specified relative configuration.
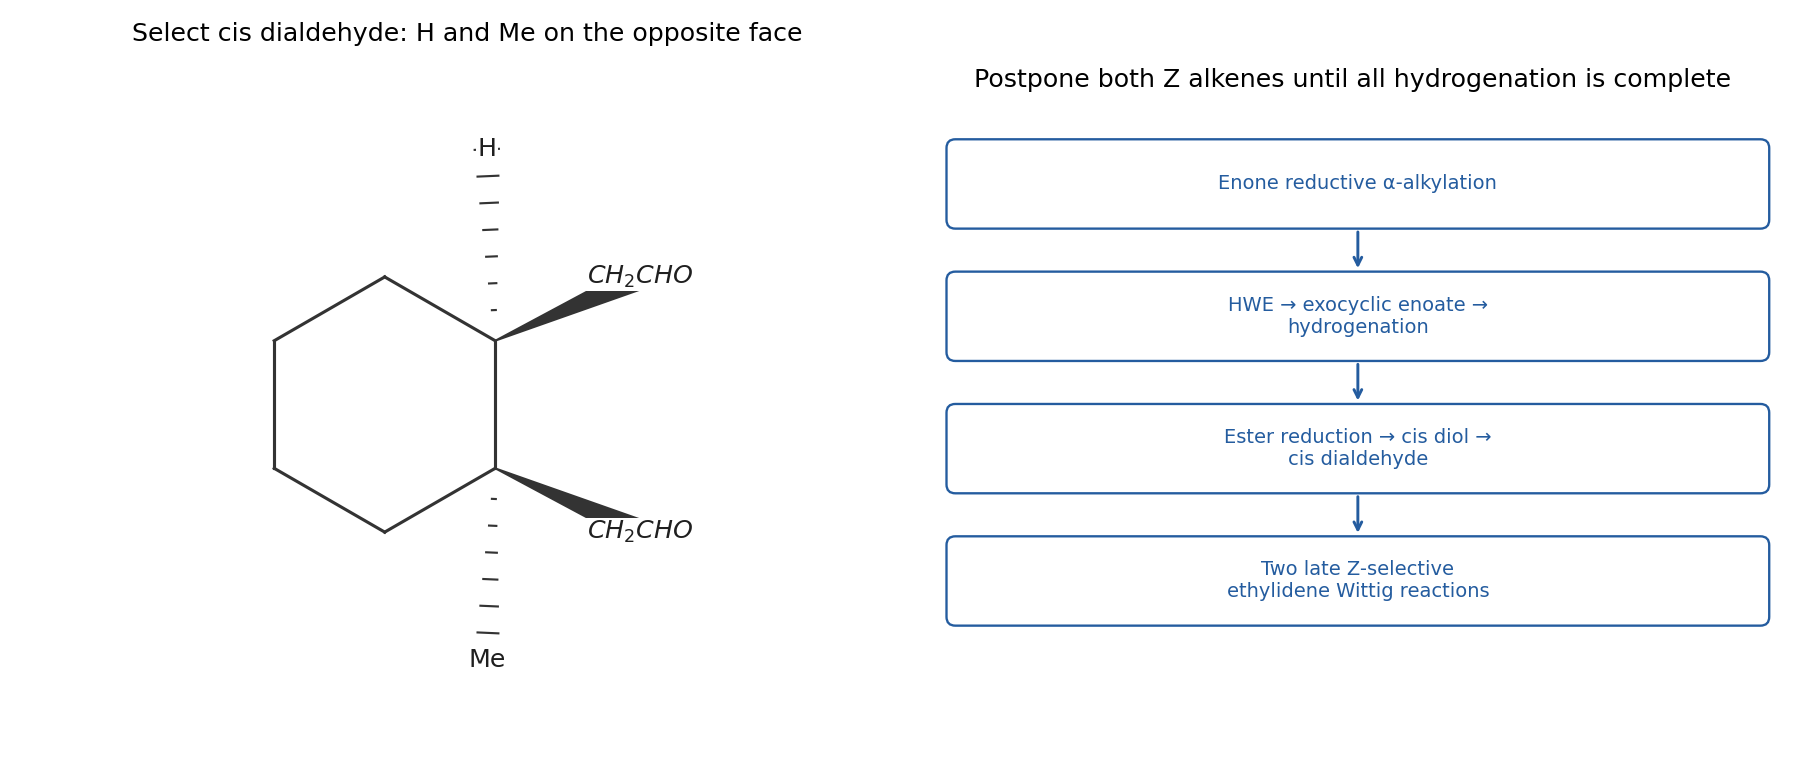
Original stereochemical plan through a cis diol/dialdehyde and late double Z-selective Wittig olefination
. Details. 
Use the nitro group as a masked carbonyl and a carbon nucleophile. The most informative disconnection reverses the six-membered aldol annulation. Before that closure, the cyclopentanone beta carbon bearsThe carbon carrying will become the lower carbonyl of the final six-membered ring by a Nef reaction. It comes from the specified nitromethane. The remote methyl ketone supplies the upper methyl-substituted alkene carbon after aldol condensation.
First, use a mild base to generate a nitromethane-derived nitronate and add it conjugatively to the indicated cyclopent-2-en-1-one. Protonation gives 3-(nitromethyl)cyclopentanone. Deprotonate its nitromethyl group and perform a second Michael addition to methyl vinyl ketone, giving . The nitro-stabilized carbon is appreciably more acidic than a simple ketone alpha carbon, permitting selective nitronate formation; control the acceptor stoichiometry to avoid a further Michael alkylation of the remaining nitro-bearing H.
Form the enolate/enamine at the cyclopentanone carbon between its carbonyl and the side-chain-bearing carbon. Intramolecular attack on the remote methyl ketone closes a six-membered ring; the two shared atoms are the original enone alpha and beta carbons. Hydrolysis if required and dehydration create the alkene between the original alpha carbon and the newly formed tertiary carbinol carbon. That carbinol carbon retains its methyl substituent, placing the methyl exactly as required. The original cyclopentanone carbonyl remains the carbonyl of the five-membered ring.
Finally, form the secondary nitronate and carry out controlled acidic Nef hydrolysis to replace its unit by . The resulting fused product contains the two required ketones. Carrying out the annulation before exposing this second ketone reduces competing enolate/aldol pathways that would be available in the corresponding triketone. The nitro intermediate's competing deprotonation means that the aldol step needs conditions designed to access the ring carbonyl enolate/enamine; simply treating every carbonyl position as equally reactive would not establish the route.
The carbon ledger is from cyclopentenone, from nitromethane and from methyl vinyl ketone, giving the nine-carbon fused skeleton plus its methyl substituent. A primary-nitro Nef reaction performed before the second carbon alkylation would instead give an aldehyde and would lose this useful protected-carbonyl construction.
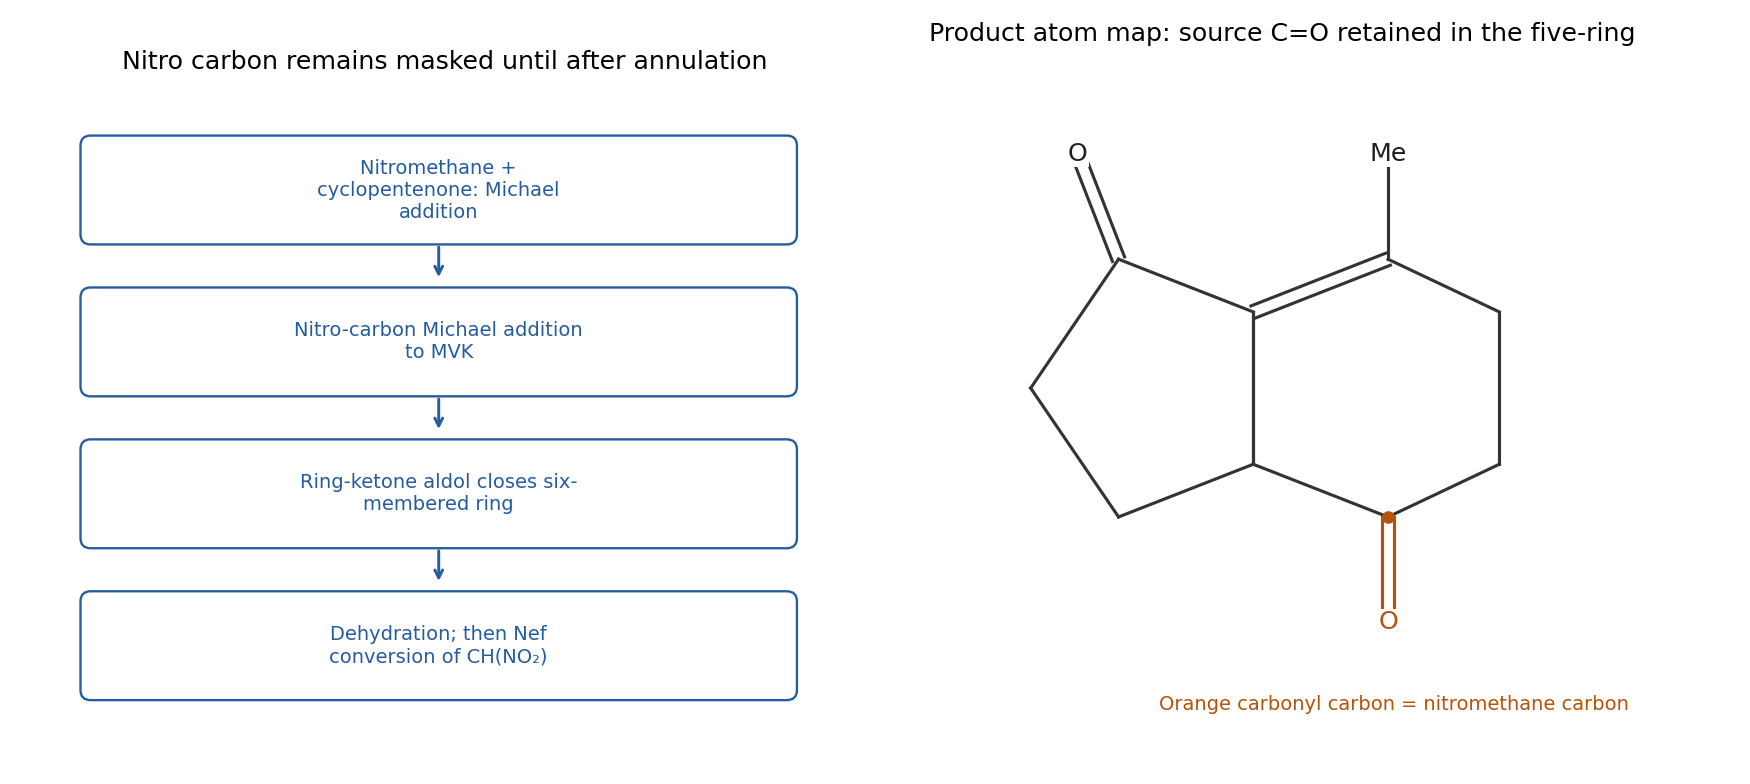
Original tandem Michael–Michael, six-membered aldol and late Nef plan, with the nitromethane carbon mapped to a target ketone
. Details. 
The left ring system is spiro, and the right-hand substituent is , not OMe directly attached to the cyclohexene ring. Let denote the right cyclohexenyl group bearing that methoxyethyl substituent cis to its linker. The connection to the left alkene carbon is . These observations suggest two Diels–Alder constructions. A left-hand dienehas the original cyclopentane carbon as one terminal diene atom; cycloaddition with an ethylene equivalent gives the spiro six-membered ring. Its residual double bond is between the two internal diene atoms, putting the linker on the required alkene carbon.
Build and its four-carbon alcohol appendage first. Protect the OH group of the indicated 4-Bromobutan-1-ol, for example as a THP ether. Convert the primary bromide to its grignard reagent and formylate with DMF, giving . olefination with an allylidenetriphenylphosphorane, , supplies a conjugated diene ; select the required diene geometry rather than assume a single E/Z outcome.
Cycloaddition with methyl vinyl ketone produces a cyclohexene with a protected four-carbon alcohol chain at one saturated ring carbon and an acetyl group at its adjacent ring carbon. Choose and isolate the adjacent-substituent cis regioisomer, matching the two dashed substituent bonds in the target. Endo preference and diene geometry can aid the selection, but an unsymmetrical diene is not guaranteed to give a unique regioisomer. Reduce the acetyl ketone to , then methylate that alcohol with sodium hydride/methyl iodide while the chain-end OH is protected. Remove the THP group under mild acid conditions to reveal , preserving the secondary methyl ether and the ring alkene.
Oxidize this primary alcohol to . Add vinylmagnesium bromide to give , then oxidize the allylic alcohol to the enone . This carbonyl carbon is the original alcohol-end carbon of 4-bromobutanol; after olefination it will be the linker-bearing carbon of the left ring alkene.
In the other branch, reduce the specified cyclopentanone to cyclopentanol, convert to bromocyclopentane, and react with triphenylphosphine to obtain the cyclopentylphosphonium salt. Deprotonation gives the cyclopentylidene phosphorane, whose ylidic carbon is part of the five-membered ring. Wittig olefination of the just-prepared enone gives the substituted diene displayed above. This carbonyl–ylide pairing keeps the ring carbon terminal in the diene and avoids accidentally constructing a fused ring instead of a spiro ring.
Finally, cycloadd an ethylene equivalent. Ethylene itself requires suitably forcing/high-pressure conditions; a more reactive alternative is phenyl vinyl sulfone, followed by reductive removal of the sulfone substituent using a method compatible with the methyl ether and retained C=C bonds. The latter installs the same two otherwise unsubstituted saturated carbons. Control or separate any additional cycloaddition isomers, and retain the previously selected cis configuration on .
The carbon count provides a strong check: cyclopentanone supplies five, bromobutanol four, DMF formylation one, the first allylidene ylide three, methyl vinyl ketone four, O-methylation one, vinylmagnesium two, and the final ethylene equivalent two, totalling 22 target carbons. Three original bromobutanol carbons form the linker; its fourth becomes the substituted carbon of the spiro ring's alkene. The proposal is a multistep constitutional/relative-stereochemical route with explicit regioisomer and diastereomer selection, not an unsupported claim of a single quantitative cycloaddition product.
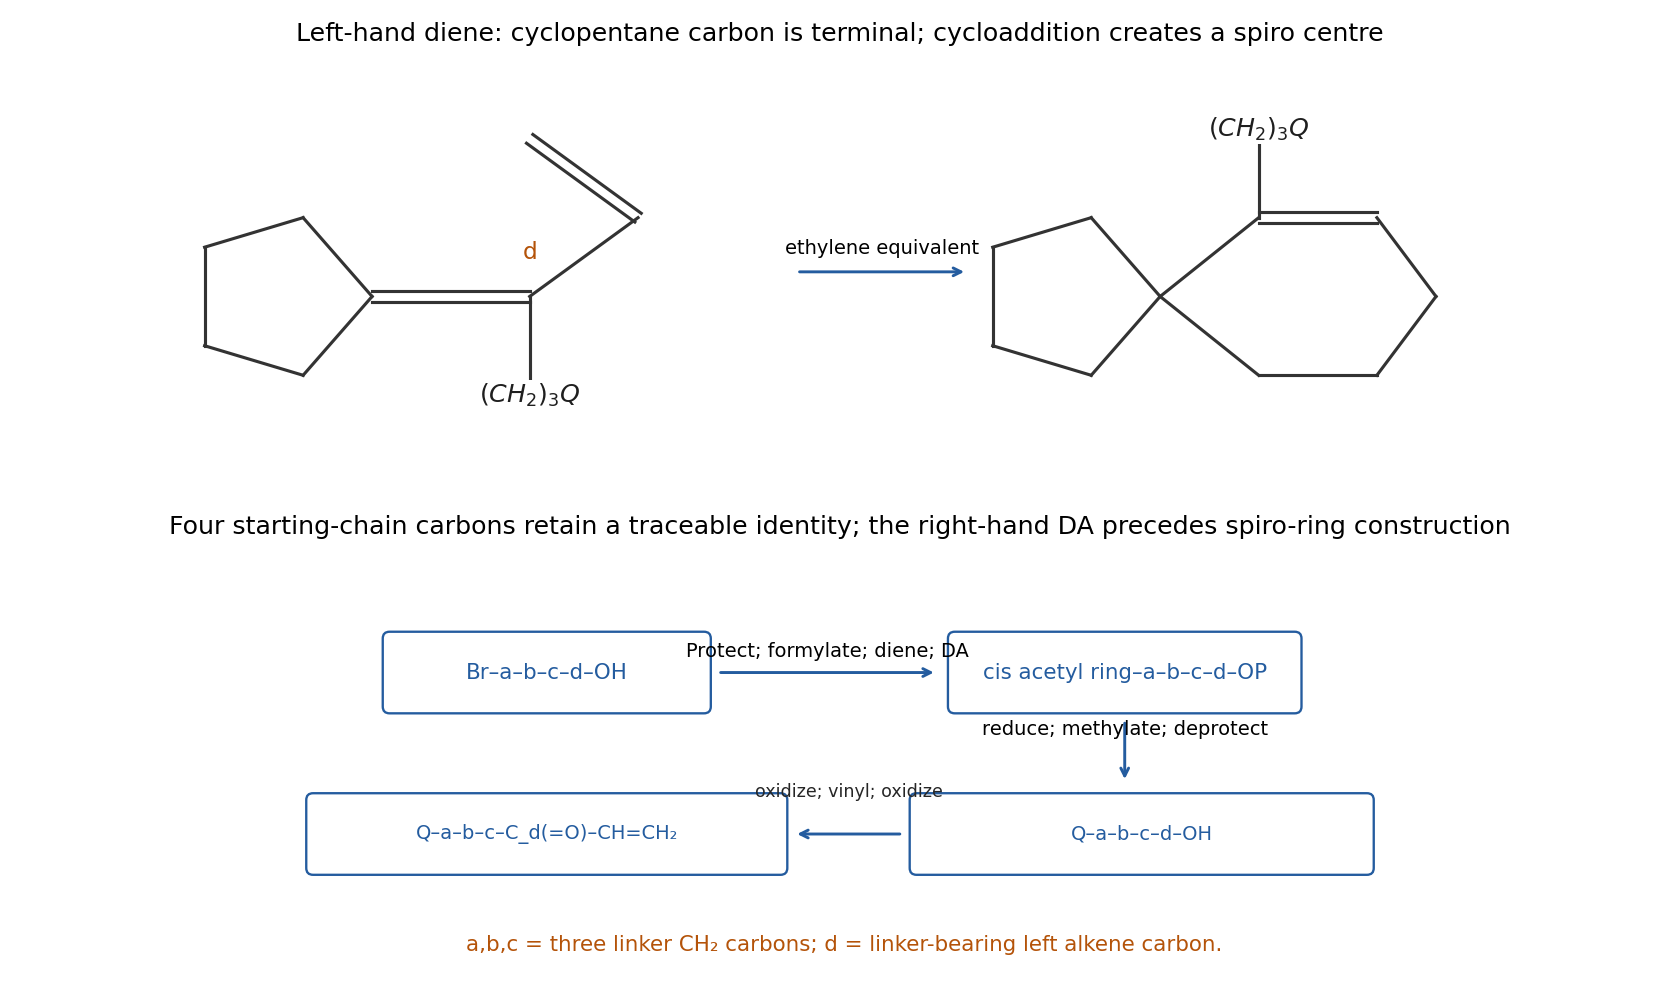
Original two-cycloaddition spiro synthesis with the four bromobutanol carbons mapped to the three-carbon linker and its adjacent alkene carbon
. Details. 
Using mass for the dominant lithium isotope gives a transition wavenumber . Inserting the thermal speed into the stated Doppler broadening expression givesThe supplied formula is the Gaussian half width at half maximum; the corresponding full width is about . Meanwhile exponential population decay with lifetime produces a natural linewidth expressed as a full widthHere must be in in the second expression. Equivalently the fine-structure separation is about GHz, the Doppler half width about GHz, and the natural linewidth expressed as a full width about MHz. The natural linewidth is therefore negligible compared with thermal broadening; low density also makes collision broadening less important.
The doublet is resolvable in principle, but only marginally against the Doppler width. The separation exceeds one Doppler full width, so two components can survive, although their profiles overlap strongly. The and upper levels have different statistical weights, so the weaker member need not make an equally conspicuous maximum. A spectral resolving power is the basic instrumental requirement; a width well below and adequate signal would make the test more convincing. An arbitrarily fine spectrometer cannot remove the physical Doppler overlap. This conclusion uses the stipulated thermal, optically thin picture; additional photospheric velocity fields or optical-depth effects would require more information.
Use the D3h molecular point group for phosphorus pentafluoride, with along the two axial P–F bonds. For each symmetry operation, only atoms left in place contribute to the character of the Cartesian displacement representation; each contributes the trace of the corresponding spatial matrix. In the class order this givesFor example a fixes P and one equatorial F, and each fixed atom has Cartesian trace , giving . reflection in the equatorial plane fixes four atoms, each with trace , giving . Reduction with the character table givesSubtract translations and rotations to obtainIts dimension is , as required for a nonlinear six-atom molecule. Thus there are eight symmetry-distinct fundamental frequencies in the harmonic picture, with each E mode doubly degenerate. This analysis concerns vibrations about one trigonal-bipyramidal minimum; it does not treat large-amplitude rearrangements between minima.
The octahedral splitting separates the metal and orbital sets. The Racah parameter characterizes interelectronic repulsion and hence the spacing of many-electron d-shell terms; it is not another name for . A Tanabe-Sugano diagram plots term excitation energies against , for specified values of the other repulsion-parameter ratios. Each ordinate is measured relative to whichever term is the ground state at that field strength. The original many-electron treatment diagonalizes the relevant d-shell energy matrices rather than identifying every absorption with a one-electron gap.[ref]
On the Absorption Spectra of Complex Ions. I by Yukito Tanabe and Satoru Sugano. 1954, pp. 753–766, d-shell energy matrices in a cubic field
The ground state changes from high-spin , , to low-spin , . The larger splitting eventually compensates the additional pairing cost. Because all plotted excitation energies are re-referenced to the new ground state, the branches change slope at the crossover. The break is not an infinite physical excitation energy or a discontinuity of the underlying Hamiltonian.
Here , so the ground term is . There are no spin-allowed d–d bands in the displayed manifold under the spin selection rule: the excited terms shown have doublet or quartet rather than sextet multiplicity. Very weak spin-forbidden absorptions can still gain intensity through spin–orbit coupling, and even spin-allowed d–d transitions would be forbidden by the Laporte rule in ideal centrosymmetric coordination. Thus a strong ordinary d–d spectrum should not be predicted simply by multiplying the splitting by a numerical factor. For scale, the lowest quartet branches read roughly at this abscissa, corresponding to weak features around , and if such forbidden intensity is detectable. These are graphical estimates, not precision assignments of every weak line.
Now and the ground term is . Spin-conserving transitions to the labelled doublet excited terms are possible. Reading the diagram gives approximatelyThe first two upper terms are coincident or unresolved at this diagram's resolution. These doublet transitions should dominate over the spin-forbidden transitions to sextet or quartet levels. Their intensity still depends on vibronic or other relaxation of the Laporte rule. The plotted curves justify only approximate wavenumbers, not several-digit band positions.
Let and treat the quencher concentration as a constant reservoir. Weak steady excitation, one emitting population, and the three specified processes giveDividing the fluorescence photon-production rate by the absorbed photon rate therefore givesThis assumes no saturation, photochemical loss, reabsorption, static association or additional excited-state decay, and that each counted absorbed photon reaches the stated population. In this restricted scheme and the unquenched lifetime is .
Rearrangement gives the Stern-Volmer equationThus a straight-line plot against has unit intercept and slope . Determine using an independently measured unquenched lifetime, or use known in the restricted model. Yield measurements alone do not separate two unknown rate constants. Equivalently the slope of is .
Use the absorbed intensity rather than incident intensity to normalize samples, and correct solvent/background signals, quencher absorption and fluorescence reabsorption. Under dynamic fluorescence quenching the lifetime ratio should follow the same straight line as the yield ratio. A reduced intensity without the corresponding lifetime change suggests static fluorescence quenching or an optical artefact; curvature can also indicate multiple populations or more complicated kinetics.
A quencher-independent intersystem-crossing rate adds to the unquenched decay rate:The normalized Stern-Volmer equation retains the same form, so intersystem crossing does not bias if the actual unquenched lifetime is used. It reduces below one and changes the slope through ; assuming would then give the wrong result. A quencher that itself changes intersystem crossing or returns triplet population to the singlet requires a larger kinetic scheme.
Writing and , the normalized determinant isIts two spin orbitals are orthogonal because , even if the spatial atomic functions have nonzero overlap. Exchanging both the spatial and spin coordinates of the two electrons reverses the sign, as required for fermionic antisymmetry.
At this separation the orbital overlap and interatomic two-electron exchange integrals are negligible. Each localized electron then contributes essentially the isolated hydrogen energy, while the interatomic electrostatic terms cancel to leading order for neutral fragments. Including nuclear repulsion in the molecular energy givesThe closed-shell RHF determinant instead doubly occupies . It retains ionic configurations with both electrons on the same atom, even at dissociation. Using atomic units and the Hartree energy in the stated unrelaxed hydrogenic 1s basis, the same-atom two-electron Coulomb integral is . Consequently at infinite separationwhere energies and integrals inside the first expression are in atomic units. It is much too high compared with two neutral atoms. Retaining the leading intersite Coulomb term at gives , still about above UHF. The numerical RHF estimates specify the supplied fixed 1s basis; a larger optimized orbital space changes them. If nuclear repulsion is excluded by an electronic-energy convention, subtract the same from both quoted total molecular energies. The restricted/unrestricted comparison and their dissociation behaviour are unchanged.[ref]
XXXIV. Notes on the molecular orbital treatment of the hydrogen molecule by Charles Alfred Coulson and Inga Fischer-Hjalmars. 1949, pp. 386–393, minimal atomic-1s molecular-orbital treatment
The fixed localized determinant is unsuitable near equilibrium. It does not develop the correct shared bonding orbital and is not a pure singlet: for one alpha and one beta orbital, , which is nonzero unless the occupied spatial orbitals coincide. At dissociation it is an equal singlet/triplet mixture, though their energies become degenerate there. Near equilibrium the physical ground state is a bonding singlet. An optimized UHF calculation can recover the restricted, spin-pure solution by making its two occupied spatial orbitals equal; that does not rescue the particular fixed atomic-orbital determinant written here. A spin-adapted multideterminant description is needed to combine good bonding near equilibrium with correct singlet dissociation.
Number the four sites of the cyclobutadienyl dianion cyclically and apply the Hückel method. The orthonormal pi basis givesA convenient real orthonormal eigenbasis isThus one bonding and two degenerate nonbonding orbitals are filled, and the antibonding orbital is empty. There are six pi electrons, including the two added by the dianion charge. Rotating the two occupied nonbonding orbitals within their degenerate subspace changes their individual sketches but not the total density matrix.
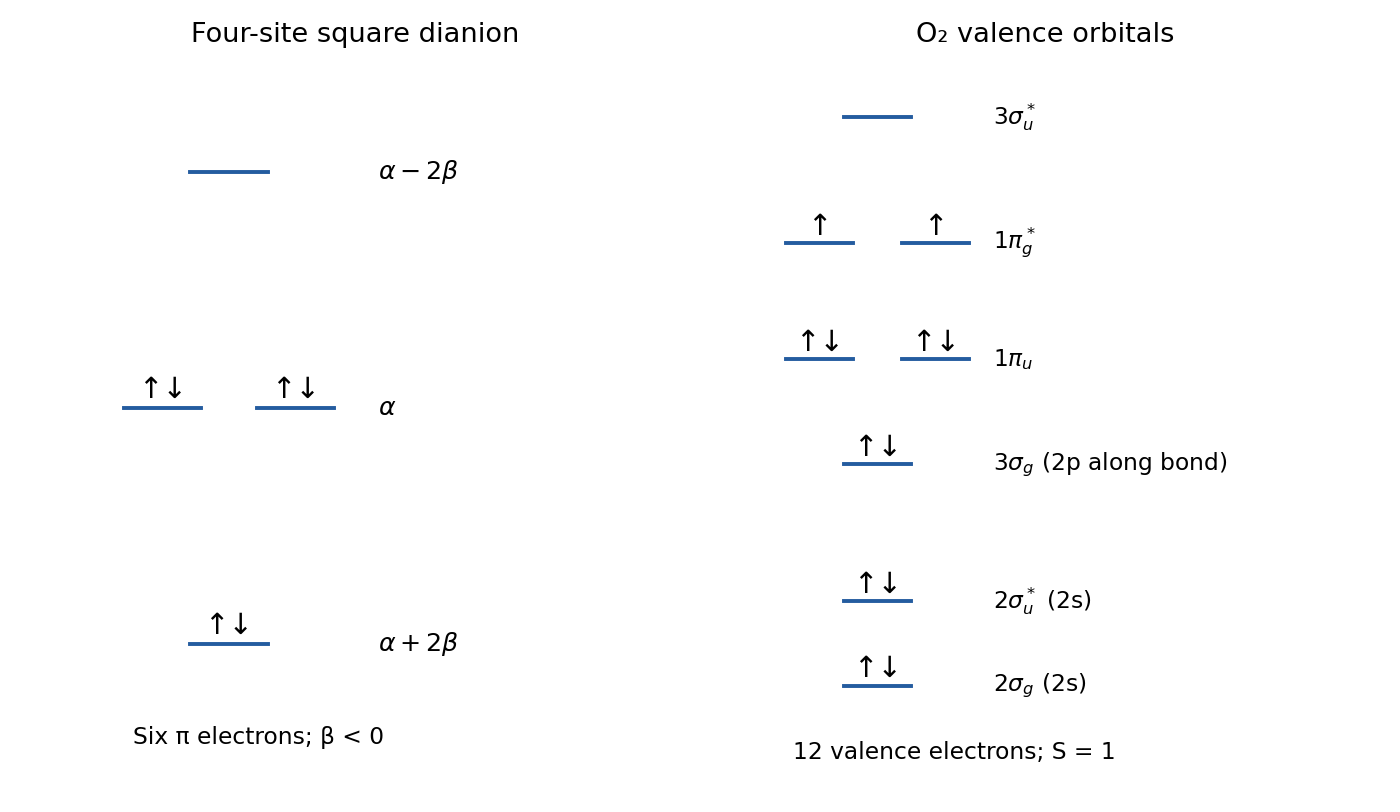
Original orbital levels and occupations for the four-site dianion and valence O2
. Details. 
The Hückel orbital charge density and Hückel pi bond order giveEach site receives electron from the lowest orbital and one from the occupied nonbonding subspace, so at every carbon, summing to six. Relative to one neutral pi electron per carbon this is a pi charge of at each site.
For adjacent sites, only the lowest orbital contributes, giving . For opposite sites the nonbonding contribution reverses the sign: . These non-neighbour negative density-matrix entries are not extra sigma bonds or extra resonance integrals in the Hamiltonian. If only skeletal nearest-neighbour pi bond orders are requested, the four values are the relevant ones.
configuration interaction with singles and doubles (CISD) uses a linear combination of the reference Slater determinant and singly and doubly excited determinants. Its many-electron trial state is therefore not restricted to one determinant; the orbitals may be held fixed while the configuration coefficients are varied.
Articles were limited to the first 100 out of 963 total.
Articles by others on the same topic
There are currently no matching articles.