For an isotropic free-electron gas at zero temperature, the Fermi wavevector is the radius of the occupied region in wavevector space. Its energy is . The first Brillouin zone is the Wigner-Seitz cell of the reciprocal lattice; it contains one distinct crystal momentum per primitive cell and per band, with spin counted separately.
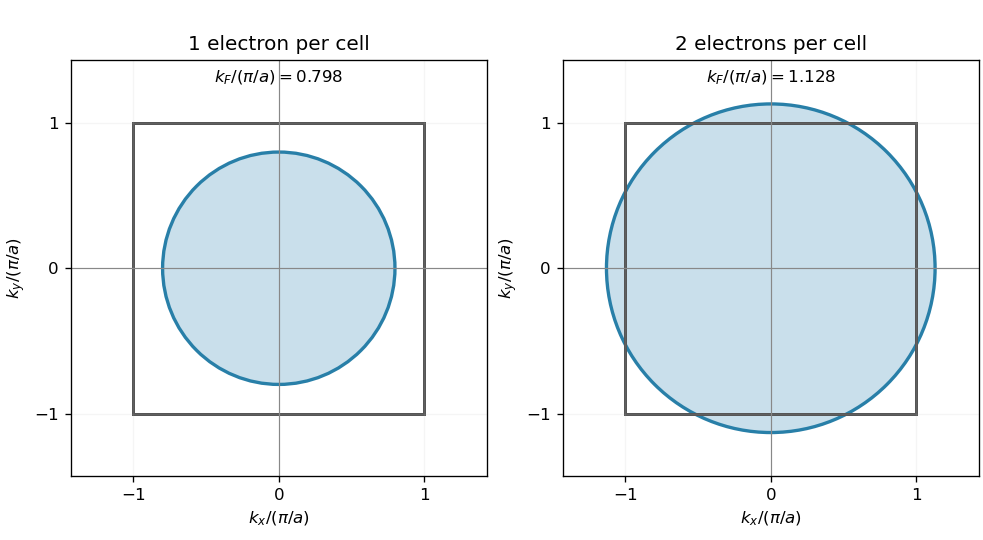
For a two-dimensional sample of area , allowed wavevectors have density . Including two spin states, filling a circle gives . Dividing by area therefore yieldsThe square lattice has one atom per area , so for valence the free-electron density is . Its Brillouin zone has . ThusThe monovalent Fermi circle lies inside the square. The divalent circle crosses the faces but misses the corners; its area equals the zone's area. These are the free-electron contours before distortion by the periodic potential.

At a Bragg plane, the periodic potential mixes degenerate plane waves differing by a reciprocal lattice vector. The two standing-wave combinations have different charge densities relative to the ions and split in energy, opening the gap. In a two-wave approximation, the boundary energies are , with .
The boundary wavevector magnitude is along and along . The free-electron boundary energies are thereforeWithin the stated direction-independent gap picture, the upper band's lowest face-edge energy is approximately , while the lower band's corner-edge energy is approximately . There is consequently a small overlap, about , rather than a global insulating gap. The divalent Fermi level lies in that overlap, producing occupied upper-band states and unfilled lower-band states.
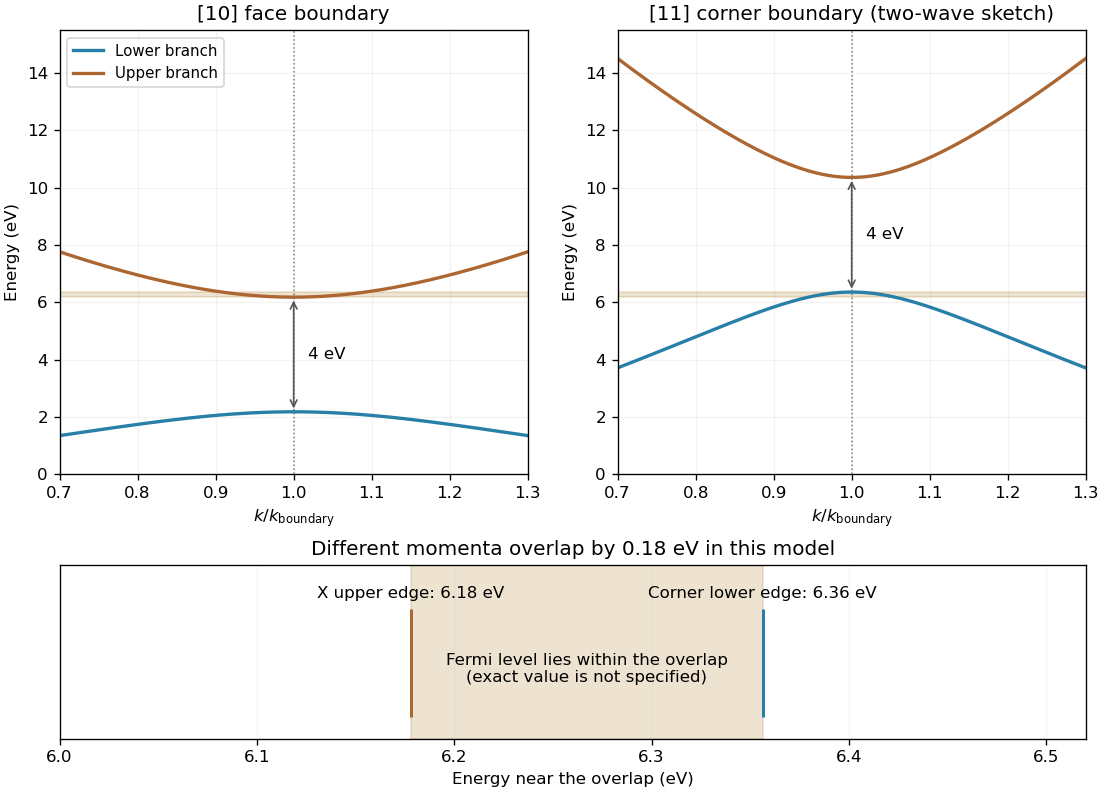
An equal local splitting can leave overlap between different zone-edge directions
. Original schematic two-wave dispersions near the and boundaries. The shaded Fermi-level range marks the overlap implied by the question's direction-independent gap approximation; no precise Fermi energy is inferred. These separate directional sketches do not specify a complete two-dimensional periodic potential. Details.
Thus the intended isotropic-gap approximation predicts a metal, with small electron and hole pockets. Positive direct gaps can coexist with band overlap. A fully specified square-lattice potential would require simultaneous treatment of the corner degeneracies; a numerical boundary splitting alone does not uniquely determine an arbitrary two-dimensional band structure. The conclusion here uses the given simplified gap picture, rather than electron count alone.
Articles by others on the same topic
There are currently no matching articles.