An isolobal analogy compares fragment frontier orbitals, not atomic sizes or total electron counts. The neutral BH fragment has four valence electrons; two make the external B–H bond, leaving two skeletal electrons in three frontier orbitals directed towards possible cluster neighbours. For the neutral fragment, contributes five electrons and cobalt nine, giving 14. Removing the 12 electrons assigned to filled, predominantly non-skeletal fragment orbitals again leaves two electrons in three cluster-directed frontier orbitals.
The corresponding frontier sets have comparable numbers, spatial symmetries and occupations, allowing replacement of a BH vertex by CpCo in a suitable cluster-electron count. This is why the metal fragment can cap an open carborane face without adding an arbitrary new set of skeletal electrons. Exact orbital energies, ligand preferences and bond lengths need not be identical.[ref]
Building Bridges Between Inorganic and Organic Chemistry (Nobel Lecture) by Roald Hoffmann. 1982, isolobal fragment correspondence and its use across organic/inorganic bonding
React pentaborane with an alkyne under relatively mild conditions to obtain a nido framework, conventionally nido-2,3-dicarbahexaborane(8) for acetylene. The two entering carbon atoms initially retain their adjacency. A specific mild example uses 2-butyne with triethylamine-assisted at or below room temperature to give nido-. This is a practical route to an explicitly nido carborane, rather than an assumption that an alkyne supplies a closo cage under every condition.[ref]
Improved Synthetic Routes to Small Carboranes: Bench-Scale Preparation of nido-2,3-C2B4H8 Derivatives from B5H9 and from B3H8- Salts by Narayan S. Hosmane and Russell N. Grimes. 1979, abstract and preparation discussion: amine-assisted substituted-alkyne reaction under mild conditions
More energetic gas-phase reaction or pyrolysis promotes hydrogen loss, boron-fragment loss and cage rearrangement, giving closo carboranes, for example - and -, with other cage sizes possible in the mixture. Thermolysis of a previously formed nido carborane is an alternative route to this stage. The point is not a clean same-composition nido-to-closo isomerization: the change of hydrogen content and sometimes cage size must be retained in the formulas.
Milder conditions favour kinetically trapped adjacent-carbon products; higher temperatures permit migration to carbon-separated arrangements, often increasing favourable B–C contacts relative to adjacent C–C/B–B contacts. Conditions and Lewis bases also influence which separated-carbon cage is isolated; temperature alone does not prescribe a single quantitative isomer distribution. Primary thermolysis studies and the small-carborane literature document both cage rearrangement and reagent-dependent product mixtures.[ref]
Synthesis of Carboranes from Dihydrocarboranes by Thomas P. Onak and others. 1963, printed p. 3378, introduction and product summary
For , twelve BH vertices supply skeletal electrons and the charge adds two:For the cobaltacarborane, nine BH vertices supply 18, two CH vertices supply six and CpCo supplies two by (a):Both are 12-vertex closo icosahedra with 13 skeletal electron pairs. In the cobalt compound, the Cp ring is external to the cage. cobalt caps the five-membered open face of the 11-vertex dicarbollide fragment and is a vertex of the resulting icosahedron, giving the usual adjacent-carbon cobaltacarborane framework. Counting all five Cp carbons as additional polyhedral vertices would be incorrect.
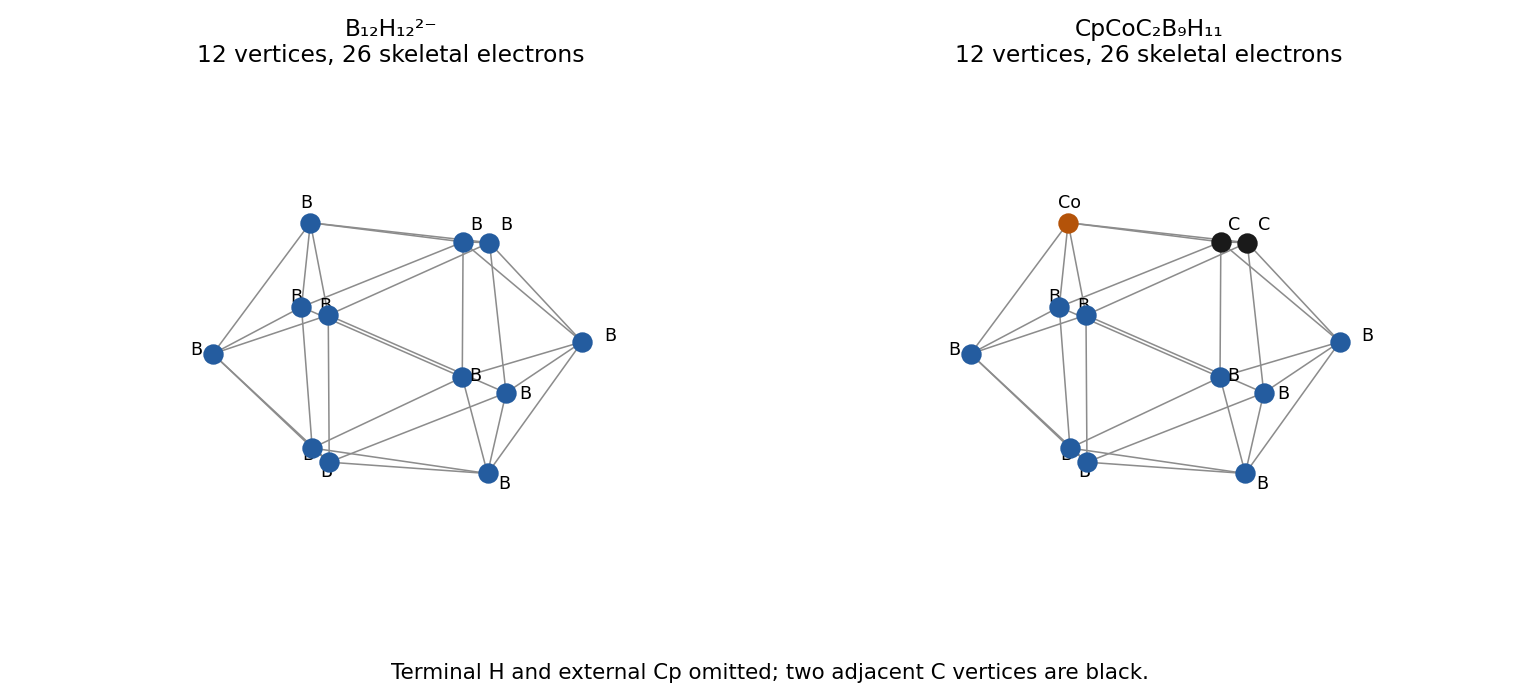
Original 12-vertex closo frameworks for dodecaborate and the adjacent-carbon cobaltacarborane, with external ligands omitted
. Details. 
Write for the open-face dicarbollide ligand. The structure consistent with the stated attachment is , with a direct Mo–Mo connection, three carbonyls on the cage-capping Mo and five on the outside Mo. The first Mo has an interaction with the five-vertex face and completes a 12-vertex metallacarborane cage; the second is exo to that cage. The two tetramethylammonium ions are counterions.
The unsaturated fragment is a 16-electron Lewis acceptor; is an 18-electron metal-base fragment in ionic counting, . Donation from a filled Mo-based orbital supplies the external fragment with a pair. This donor–acceptor representation avoids the mistaken claim that both initially saturated 18-electron fragments simply add a new conventional bond without changing anything.
An alternative formal oxidation-state allocation for a single covalent Mo–Mo bond is cage Mo(I) and external Mo(−I): the local counts are and . These are electron-bookkeeping alternatives to the metal-base/metal-acid picture, not a measurement of integral site charges. With terminal carbonyls, the external group can be approximately octahedral when the Mo–Mo direction occupies its sixth coordination direction. The cage-containing unit must accommodate an exo metal interaction in addition to its facial cage/CO environment; its geometry need not be a perfectly undistorted isolated half-sandwich.
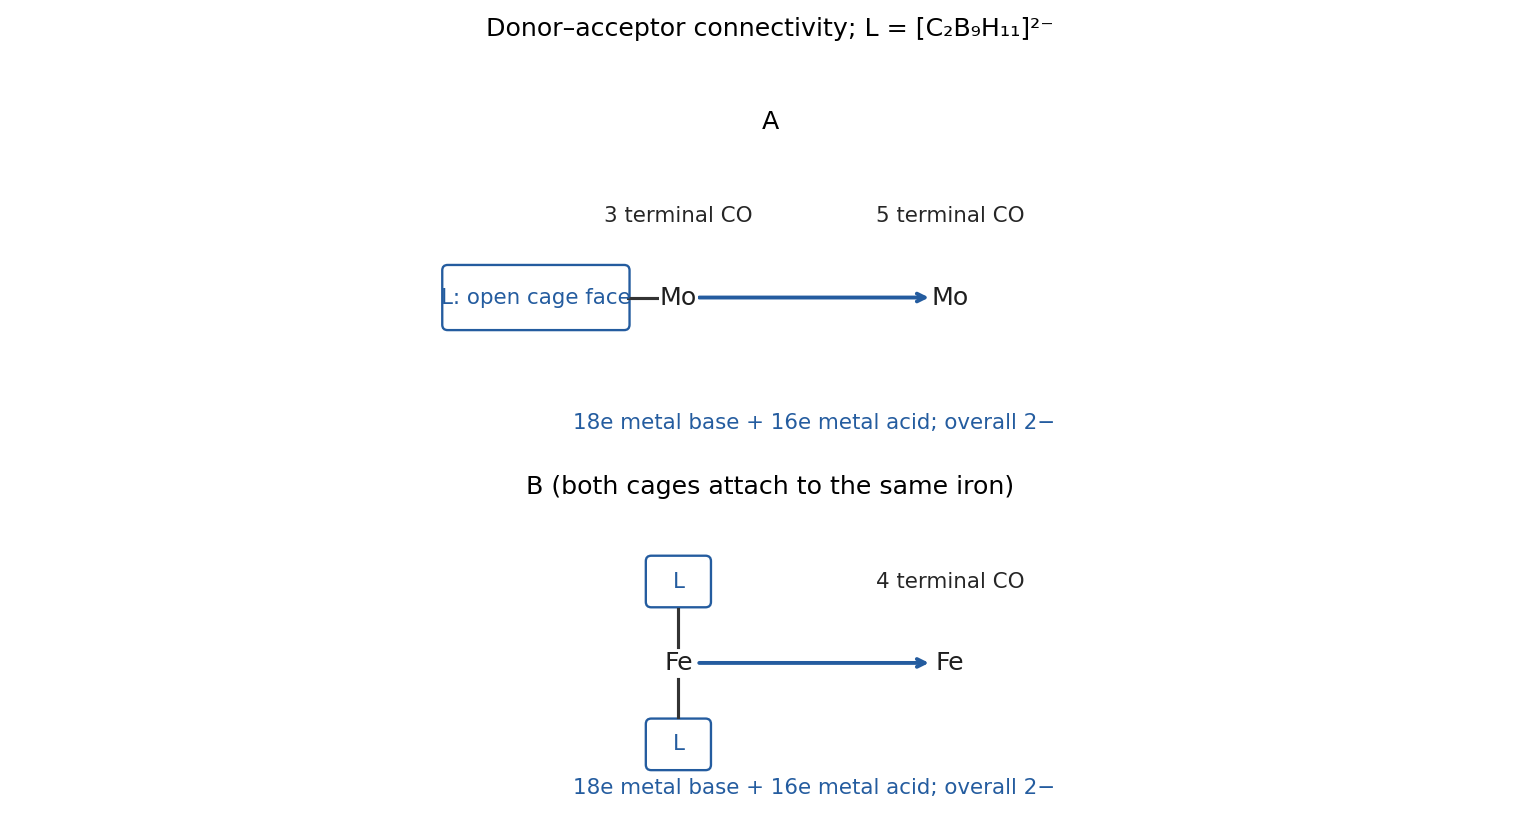
Original donor–acceptor models for the exo Mo and Fe carbonyl fragments attached to cage-bound metal donors
. Details. 
The wording places both dicarbollide cages on the same iron, leaving the second iron outside the cages. The corresponding proposal is , with the first iron sandwiched between the two open faces and the second carrying four carbonyls. It is not the symmetric arrangement with one cage on each iron, which would contradict the specified single cage-interacting metal site.
The donor fragment has electrons; the external neutral fragment has . Metal-to-metal donation therefore provides a natural 18-electron acceptor adduct. In covalent-single-bond oxidation-state bookkeeping, the analogous assignments Fe(III)/Fe(−I) give and . In either account the whole species has two negative charges. The outside iron can have an approximately trigonal-bipyramidal five-coordinate arrangement when the Fe–Fe direction and its four CO ligands are counted.
The important structural distinction is a bis(dicarbollide) iron donor joined to an exo tetracarbonyl iron acceptor. Formula and electron counting alone do not uniquely fix all carbonyl angles or prove a particular crystalline conformation.
Neither formula requires a bridging carbonyl to repair an electron deficiency: the terminal-CO donor–acceptor structures in (d) and (e) already give an electron-counting rationale for the metal–metal interaction. A predominantly terminal-carbonyl model is a reasonable starting proposal for A; bridging or semibridging CO is more geometrically plausible for the shorter Fe–Fe contact in B. Larger metal separation makes a carbonyl simultaneously engaging both Mo atoms less favourable, while carbonyl redistribution or tilting can be easier around a compact Fe pair.
This is a qualified structural prediction, not a determination from molecular formula. A bridging CO can change the relative hapticity and local electron allocation, and the two metal centres are inequivalent; a large frequency shift by itself need not prove a symmetric bridge. Infrared carbonyl frequencies combined with diffraction would test terminal versus bridging or semibridging assignments. The given low-temperature preparation can trap a coordination pattern before complete carbonyl redistribution.
Articles by others on the same topic
There are currently no matching articles.