Make heterogeneous electron transfer fast relative to delivery of reactant, usually by applying sufficient overpotential so that its surface concentration approaches zero. Use excess supporting electrolyte to suppress migration. For purely diffusional transient transport, suppress uncontrolled convection; for a rotating disk electrode, controlled convection supplies the bulk while diffusion across the thin interfacial boundary layer limits the current. A limiting current that no longer increases with driving potential is the characteristic test; a large current alone is not sufficient.
The midpoint potential characterizes the redox couple under specified solution conditions: it is the potential at which oxidized and reduced forms have equal activities, or equal concentrations in the corresponding formal-potential convention. For a reversible cyclic voltammogram it is estimated by . It approximates the formal redox potential when the usual reversibility and diffusion assumptions apply; it is not the potential halfway along a scan chosen arbitrarily.
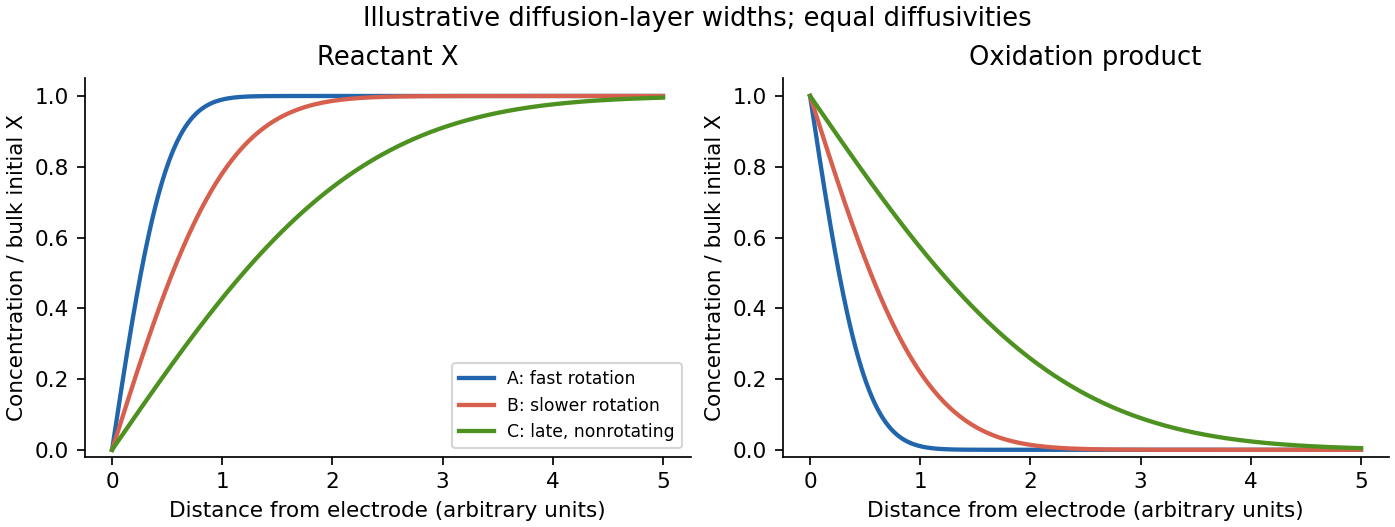
All three marked points are at strongly oxidizing potentials, where X is depleted at the electrode: . Assuming only X initially in bulk, and . Oxidation produces X at the interface.
- A: the largest steady limiting current corresponds to the fastest rotation and thinnest diffusion boundary layer. X rises steeply from zero to ; X falls steeply from its surface value to zero.
- B: slower rotation gives a thicker layer and shallower gradients, hence a smaller limiting current.
- C: the nonrotating electrode gives the peaked, subsequently decaying current. Its diffusion layer continues to broaden with time, approximately as , giving the broadest and shallowest profiles at the indicated late-scan point.
For equal diffusion coefficients, the two concentration profiles are complementary, . More generally equality of their interfacial molar fluxes requires , so the product surface concentration need not equal . A and B are approximately steady hydrodynamic profiles; C is a time-dependent diffusion profile.

Read the potential labels rather than the alphabetical order: b, the less negative event, is ; a, the more negative event, is . These are the first and second reductions respectively, with the reverse branches giving the corresponding oxidations. The second electron is harder to add to an already negatively charged species.
Event a probes electron transfer between the semiquinone and dianion. Benzoquinone's smaller framework localizes charge and undergoes a greater change in bonding and solvation on this reduction; a larger effective reorganization barrier can make its heterogeneous transfer insufficiently fast for Nernst equilibrium on the time scale. Anthraquinone's extended framework distributes the added charge and can support the more reversible response seen here.
This is a kinetic interpretation of quasi-reversibility, not a claim that benzoquinone cannot be reduced. Follow-up chemistry or adsorption can also distort peak separation and return-current ratios; scan-rate-dependent measurements would distinguish them from slow electron transfer. A single trace does not uniquely identify the cause.
A useful electrode-surface scheme for the proposed ECEC mechanism isElectron-transfer arrows occur at the electrode, whereas both capture steps occur in the adjacent solution layer. CO is attacked by an oxygen-centered nucleophilic component of the reduced quinone, forming an O-carbonate linkage rather than simply physically adsorbing on neutral AQ.
At potentials more negative than , direct second reduction of any unbound semiquinone is also accessible: , followed by carbonate formation. Thus the fully reduced unbound and mono-adduct species can feed the bis-adduct, and the drawing should include CO transport into the layer and adduct transport away. ECEC is the proposed sequence; competing EECC transfer/capture ordering is possible at such negative potentials.
Peak I is the cathodic reduction shifted positive of the unbound AQ/AQ event, consistent with reduction of the CO-bound semiquinone to mono-carbonate dianion. Peak II is the corresponding relatively negative-potential return oxidation of mono-carbonate dianion. Peak III, appearing farther positive on the faster return scan, is consistent with direct oxidation of the more strongly protected bis-carbonate dianion, followed by CO release.
This assignment is consistent with binding making the reduced species harder to oxidize. At slow scan, chemical interconversion and CO dissociation can feed the easier mono-carbonate oxidation pathway; at fast scan a larger fraction of kinetically retained bis-adduct survives to its direct oxidation potential. The two separated reductions also show that initial capture and its associated potential shift are not sufficient to make every ECEC step a single equilibrated two-electron wave.
These traces support coupled electron-transfer/binding chemistry but do not determine every microscopic rate constant or exclude EECC pathways. The peak assignments are species-family assignments within the stated mechanism, rather than proof of a unique elementary sequence from voltammetry alone.
Within the stated ECEC model and with CO genuinely in large excess, its concentration is effectively constant. Both capture steps are then pseudo-first-order in the quinone species, and at the same scan rate the faradaic current is linear in bulk AQ concentration.
The sketch therefore retains the peak potentials, sweep directions and relative features of the trace, including the positive-potential return feature III, but reduces the faradaic current scale by a factor of ten. Charging/background current need not scale with AQ concentration and should be shown separately. A changed normalized shape would indicate effects outside this simple scaling limit, such as CO depletion, adsorption or reactions between quinone species.
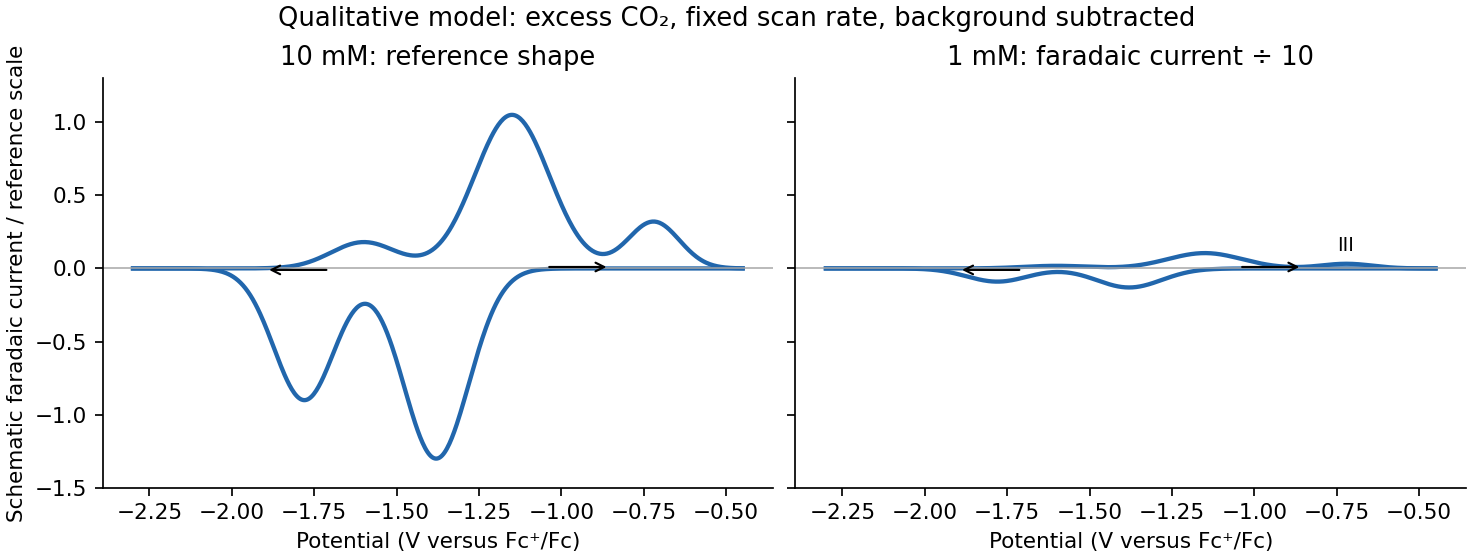
Qualitative tenfold faradaic-current scaling at fixed scan rate and excess carbon dioxide
. Details. 
The target dianion consumes two electrons per molecule, sogiving Faradaic yield. Of the passed electron equivalents, are not accounted for by the quantified bis-adduct.
They can reside in incompletely captured reduced quinones, other reduction products or parasitic solvent/proton/CO chemistry. The data do not identify one unique loss pathway. The starting solution contains BQ, so the target product represents only of that inventory; low bulk conversion alone does not explain a sub-unity Faradaic yield, because unreacted BQ has consumed no charge.
Articles by others on the same topic
There are currently no matching articles.