A route from the supplied cyclohexenone starts with conjugate methyl addition. Form an organocuprate from methylmagnesium bromide and Cu(I), or use catalytic Cu(I) with controlled methylmagnesium addition, to obtain 3-methylcyclohexanone after protonation. copper directs 1,4 rather than direct 1,2 addition, so the ketone remains intact.
Generate its less hindered kinetic enolate at the alpha carbon opposite the methyl-bearing side, using LDA at low temperature, and trap with phenylselenyl chloride. oxidation of the resulting alpha-selenyl ketone with hydrogen peroxide produces a selenoxide, whose syn elimination introduces the enone double bond on that side. Renumbering the double bond from the carbonyl gives 5-methylcyclohex-2-en-1-one A. The selenium-containing phenyl group is a removable reagent fragment, not a carbon building block incorporated into A; the only new product carbon is the allowed methyl group. Regioselective kinetic enolization is important: enolizing the other alpha site would regenerate an enone with the methyl in the wrong position. This route produces racemic A unless a separate asymmetric step is introduced, which is sufficient for the stereochemically unspecified starting ketone.
Tin tetrachloride coordinates the enone carbonyl, lowering the dienophile pi-star energy. Butadiene reacts in an endo-oriented Diels–Alder cycloaddition: its terminal carbons bond to the enone's two alkene carbons, its central bond becomes the new cyclohexene double bond, and the enone C=C pi electrons become a new sigma bond. All three bond reorganizations occur in a suprafacial six-electron cycle.
Approach from the face opposite the methyl substituent avoids steric crowding and gives the indicated relationship between methyl and the non-enolizable junction hydrogen. The immediate stereospecific cycloaddition gives a cis-fused junction relationship. The junction carbon next to the ketone can subsequently enolize; reprotonation on either face epimerizes this centre without breaking either newly formed C–C bond. This accounts for the mixture at the wavy-bond position while the other junction and methyl relationship remains fixed. The modest 1.5:1 ratio should not be mistaken for failure of suprafaciality in the cycloaddition itself. Later oxime formation removes the ketone's ordinary enolization pathway and furnishes the depicted isolated junction diastereomer.
The oxime is reduced at its C=N bond rather than cleaving N–O: hydride addition to the imine carbon followed by protonation gives an N-substituted hydroxylamine. In a chair-like model of the cis-fused ring system, the fused framework and methyl group make the two faces inequivalent. The favoured hydride trajectory accesses the face that places the newly installed C–H bond behind the original drawing and the retained C–NHOH bond in front, giving D. It avoids the more congested trajectory into the folded bicyclic framework; a flat hexagon comparison alone misses this facial distinction.
The existing methyl and junction centres are retained. The selectivity is therefore diastereoselective substrate control, not enantioselectivity from sodium cyanoborohydride, which is achiral. For a racemic starting bicycle the same relative stereochemical preference gives a racemic pair of D enantiomers.
The secondary hydroxylamine condenses with formaldehyde. nitrogen attacks the carbonyl carbon, giving a carbinolamine; proton transfer and loss of water generate the tethered nitrone , where is the entire bicyclic substituent attached to nitrogen in D. Molecular sieves remove water, favouring this condensation.
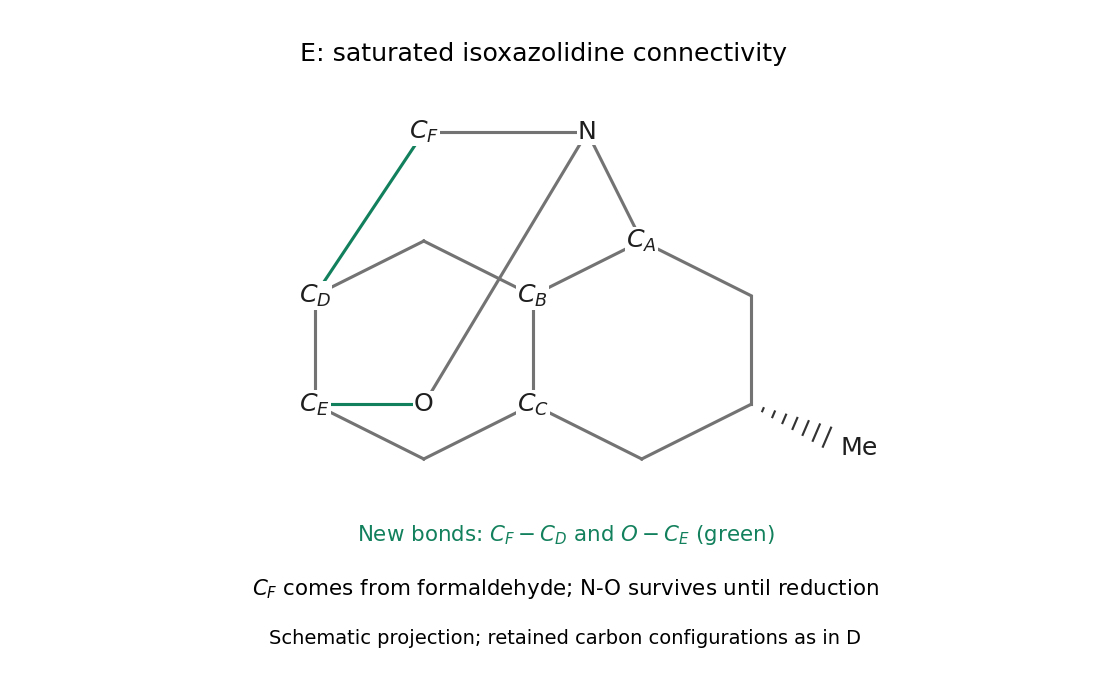
The nitrone undergoes an intramolecular 1,3-dipolar cycloaddition with the retained cyclohexene double bond. To define E precisely, label the former oxime carbon , the adjacent upper ring junction , the other junction , and the alkene carbons and such that the chain from to the more remote alkene terminus is . Let be the new formaldehyde-derived nitrone methylene. The cycloaddition forms and O–, retains N–O and N–, and changes N= and into single bonds. The new five-membered heterocycle is N––––O. These connections, together with the original bicycle, specify the bridged isoxazolidine E.
The inwardly tethered cycloaddition is suprafacial at the alkene and preserves the existing carbon stereocentres; the C–C and C–O additions occur on the same accessible alkene face. The pre-existing N-substituent orientation restricts which fold can bring both reacting ends together without crossing the bridge. This is a genuine two-new-bond cycloaddition, not a Mannich reaction followed by unrelated oxygenation.

methyl fluorosulfonate alkylates the tertiary isoxazolidine nitrogen, giving a quaternary N-methyl ammonium salt; the methyl remains attached to nitrogen in F. hydride from lithium aluminium hydride then cleaves the N–O bond, releasing the corresponding oxygen as an aluminium-bound alkoxide while restoring neutral tertiary amine nitrogen. Aqueous workup protonates the alkoxide to give the alcohol.
Thus the C–O bond made in the cycloaddition survives as the alcohol-bearing C–O bond, and the formaldehyde-derived methylene survives as the extra carbon between nitrogen and the carbon framework. The N–O bond is the one removed. N-methylation followed by reductive N–O cleavage gives the bridged amino alcohol F without requiring cleavage of its newly formed C–C bond or hydrogenation of a nonexistent residual alkene.
Articles by others on the same topic
There are currently no matching articles.