Place the trigonal axis along , the axial ligands on and the equatorial ligands in the plane. The ordering is . The pair has nodal behaviour along both the equatorial ligand directions and the axial axis, minimizing direct ligand repulsion. The in-plane pair points into the equatorial region. The lobes point at both axial ligands and its torus also samples the equatorial ligands, giving the largest repulsion in the regular point-charge geometry.
Rotation by 120° and reflection in the equatorial plane preserve the geometry; they enforce the indicated doublet degeneracies. The ordering concerns a regular trigonal bipyramid with equivalent ligand-field assumptions, not every distorted five-coordinate complex.
For two independent identical ions at sufficiently high temperature, Curie's law gives . In the cgs units used here, .
Fe(II), Co(II) and Ni(II) have , and 1, respectively. The corresponding high-spin occupancies, ordered as , areHund filling puts one electron in each orbital before additional pairing. Spin-only dimer values at would be 6.0, 3.75 and 2.0; the observed larger values anticipate orbital contributions. The spin assignments require the chemically appropriate high-spin states, rather than rounding the measured directly to a spin-only integer.
Using the spin values above and accounting for two metal ions per formula unit,The deviations from 2 indicate spin-orbit coupling and incomplete quenching of orbital angular momentum. The partly filled manifold in Fe and manifold in Ni permit orbital degeneracy in the ideal geometry. Co's high-spin orbital occupation is more orbitally balanced, but mixing with excited ligand-field states still produces a substantial effective shift. These fitted scalar values summarize a thermally averaged response; they do not determine the individual components of an anisotropic g tensor.
The radical's magnetic orbital has symmetry. Fe(II)'s partly occupied set supplies a matching metal magnetic orbital, permitting electron transfer and predominantly antiferromagnetic metal–radical exchange. In Co(II) and Ni(II), that set is filled; their unpaired metal electrons occupy orbitals of different symmetry. Approximate orthogonality suppresses the usual antiferromagnetic kinetic-exchange pathway, favouring ferromagnetic metal–radical coupling through the remaining exchange contributions.
This orbital-symmetry argument assumes the intended idealized binding geometry. It predicts exchange signs, not exact magnitudes; distortion and additional orbital mixing can alter the competition.
Adopt , with and ligand spin . Let and . The angular momentum addition identities giveThe five allowed multiplets areTheir degeneracies sum to 18, matching . For dominant ferromagnetic , the ground multiplet has , . A weak antiferromagnetic does not overcome the two strong metal–radical bonds.
A frustrated triangle has two ferromagnetic metal–radical bonds and an antiferromagnetic metal–metal bond. With the Hamiltonian convention above, take K and K. In the maximally aligned ground state the two metal–radical products are and the metal–metal product is . ThereforeHere denotes complete satisfaction of all exchange interactions; the small deviation quantifies the weak frustrated contribution in this normalization. The numerical answer assumes antiferromagnetic metal–metal exchange. The printed magnitudes alone do not establish that sign. If the metal–metal exchange were instead ferromagnetic, all three bonds would be satisfied and . The ground-state scalar products also follow directly from , , so a separate classical approximation is unnecessary for this fully polarized multiplet.
For an axial spin Hamiltonian with K, the minima are . The highest levels along the reversal ladder are the doublet. Henceabout 1.25 kJ mol or 12.9 meV per molecule. Using would incorrectly put a half-integer spin at a nonexistent level. Transverse anisotropy and quantum tunnelling of magnetization can create faster relaxation pathways, so this axial ladder barrier need not equal an experimentally fitted effective barrier.
Frequency-dependent ac susceptibility probes relaxation: a peak in the out-of-phase component occurs approximately when . Measuring its displacement with frequency determines ; an activated region may fit . A low-temperature departure can reveal tunnelling or other relaxation channels.
Low-temperature, sweep-rate-dependent magnetization hysteresis tests whether reversal is slow on the measurement timescale. A loop opening below the blocking temperature, variation with sweep rate and possible tunnelling steps provide complementary evidence. Isothermal remanent magnetization decay after field removal directly measures the associated relaxation time. A single open loop alone would not distinguish isolated-molecule blocking from collective ordering or impurities.
The framework is a three-dimensional network of corner-connected octahedra, with each apparent corner connection extended through a bridging formate, . Each metal is coordinated by six oxygen donors from six formates, which link it to six neighbouring metals in the perovskite-like topology. The guanidinium cation occupies the larger cavity and can hydrogen-bond to formate oxygen atoms. Charge balance is .
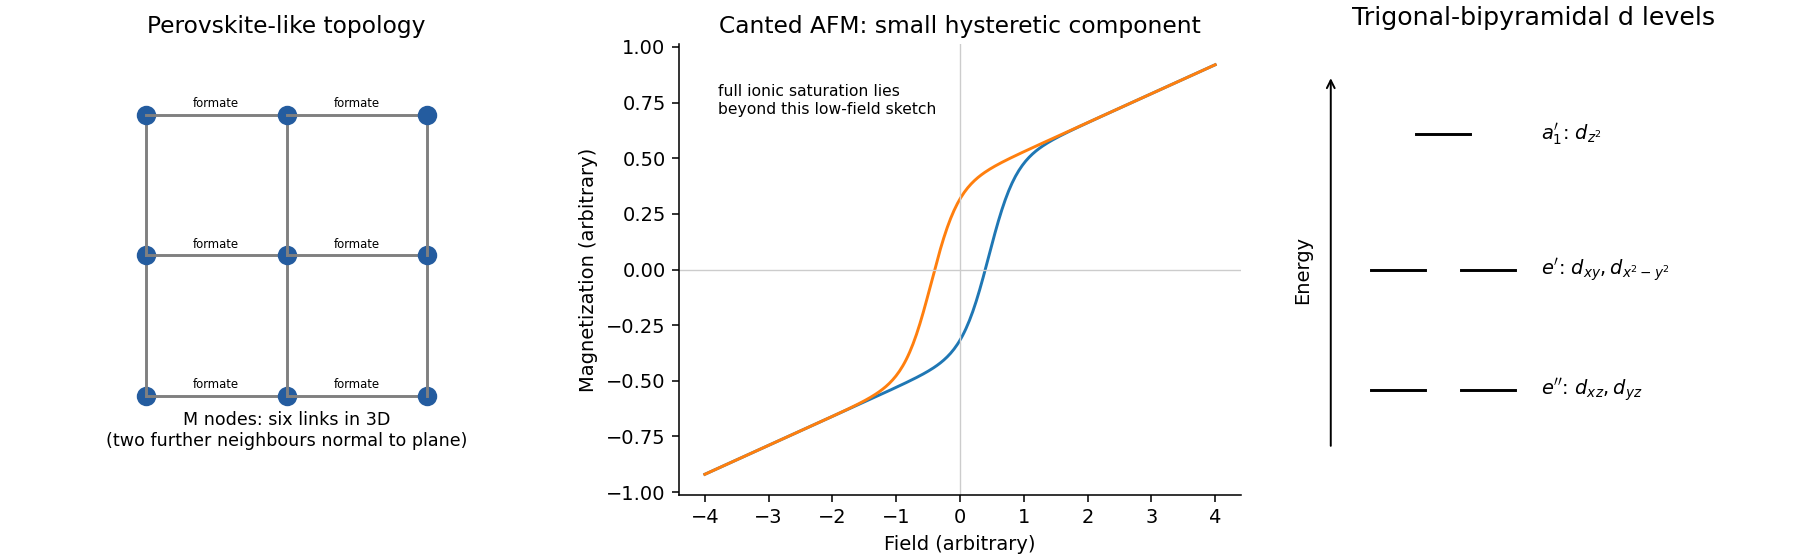
Original formate-network topology and weak-ferromagnetic magnetization schematic
. Details. 
At high temperature, approximately independent paramagnetic ions give the appropriate Curie constant. The negative Weiss temperatures and the reduction of on cooling initially indicate predominantly antiferromagnetic correlations. The sharp rise near the ordering temperature nevertheless reveals a strongly enhanced low-field magnetic response.
A net moment could arise from a canted antiferromagnet, a ferrimagnet, or a ferromagnet with competing antiferromagnetic interactions. Differences between field-cooled magnetization and zero-field-cooled magnetization suggest history-dependent domain selection or slow relaxation, but are not alone proof of a particular microscopic spin arrangement. An ordinary compensated antiferromagnet would not account for a large ordering-related ferromagnetic-like low-field contribution without an additional mechanism.
Weak ferromagnetism from canted antiferromagnetic order is the consistent model. Importantly, the plot has different magnetization scales: the left axis belongs to Mn and Fe; the right belongs to Co and Ni. Approximate low-temperature field-cooled readings are 40, 50, 450 and 390 G cm mol, respectively. Dividing by 5585 givesThese moments are tiny compared with fully aligned moments of about 5.09, 4.40, 4.25 and 2.47 , calculated using the Curie-derived values below. A conventional ferromagnet of the full high-spin ions would have much larger spontaneous magnetization; a simple bipartite array of identical metal sites also supplies no obvious unequal sublattice moments for ferrimagnetism. Opposed large sublattice moments with a small transverse uncompensated component reconcile the net response with negative .
If the 10 Oe field has aligned the weak-ferromagnetic domains, estimates sublattice canting angles of roughly 0.08°, 0.12°, 1.1° and 1.6°. These are illustrative, not precision structure determinations: the low field may not fully align the domains, and the readings are graphical. Bulk magnetization strongly supports this model but does not uniquely establish every spin direction; magnetic neutron diffraction would test the microscopic arrangement.
Use for one ion per formula unit, with high-spin for Mn, Fe, Co, Ni. ThenHigh-spin Mn(II), with an approximately orbitally nondegenerate ground state, stays closest to the free-electron value. Fe, Co and Ni have stronger orbital admixture, particularly Co in this effective Curie-law comparison. Spin-orbit coupling creates magnetic anisotropy, which can make reversal of the weak-ferromagnetic domains slower and enhance field-cooled/zero-field-cooled separation. A field-cooled sample selects domains as it orders; a zero-field-cooled sample initially has competing domains and can remain pinned when a small measuring field is applied.
This is a qualitative rationalization, not a strict proportionality between and the magnetization separation. The anisotropy tensor, domain-wall pinning and experimental timescale also matter; scalar values alone do not quantitatively fix the divergence.
Define . Each site has exchange-connected neighbours. In mean-field theory, replacing neighbours by their mean spin produces the effective field that yieldsThusfor Mn, Fe, Co and Ni. Negative means antiferromagnetic exchange in this convention; a Hamiltonian written with instead of would have twice these numerical values. The increasingly negative values indicate a growing effective exchange energy, although different orbital occupancy and anisotropy mean that they are not a simple universal function of -electron number.
A three-dimensional six-connected framework is more favourable to mean-field reasoning than isolated chains, but the theory neglects short-range correlations, anisotropic exchange and fluctuations. The ratios show substantial departures from the simplest unfrustrated isotropic mean-field relation. The values are therefore effective high-temperature exchange estimates, not a complete microscopic fit to the ordered phases.
For Co, the full field-polarized ionic value isThis is distinct from the much smaller weak-ferromagnetic domain moment, approximately 450 G cm mol at low field. A low-temperature isothermal loop should have a small hysteretic central component superposed on the antiferromagnetic field response. Reversal changes the selected weak-ferromagnetic domain and produces remanence and coercivity. At larger fields the magnetization continues to increase as the opposed sublattice moments rotate; a spin-flop or spin-reorientation feature may occur depending on field direction and anisotropy. Only sufficiently strong fields approach the full ionic saturation value, so the small low-field loop must not be labelled as saturation at 4.25 .
Below the ordering temperature, coherent spin correlations produce magnetic Bragg intensity. Depending on the magnetic propagation vector, this can appear as additional peaks or as added intensity at positions already occupied by nuclear reflections. A magnetic unit cell larger than the chemical one gives extra reciprocal-space positions; a zero-propagation-vector canted structure need not do so.
Magnetic intensity generally decreases at large scattering vector through the magnetic form factor and depends on moment components perpendicular to the scattering vector. Above the transition the long-range magnetic Bragg contribution disappears, though short-range correlations can leave diffuse scattering. The nuclear pattern remains approximately unchanged unless magnetic ordering also drives a structural distortion.
Articles by others on the same topic
There are currently no matching articles.