Both free primary allylic alcohols undergo Sharpless asymmetric epoxidation. With slightly more than two equivalents of hydroperoxide, the expected fully oxidized product is the diepoxide, with both original C=C bonds converted to epoxides and both primary alcohols retained. The two central pre-existing stereocentres and their ether substituents are retained.
The stereochemical construction is unambiguous by applying the L-(+)-tartrate mnemonic separately at each end: orient that alkene horizontally with its CHOH group at lower right, and deliver oxygen from the back face. In the source's original in-plane orientation this gives the left epoxide oxygen behind the plane and the right epoxide oxygen in front. The methyl and hydroxymethyl groups retain their original relative arrangement about each former double bond. One cannot simply draw both oxygens on the same face of the original extended chain: the two allylic-alcohol ends have opposite drawing orientations. titanium binding to each free hydroxyl directs oxygen transfer; the pre-existing stereocentres can create matched/mismatched rate differences, but the stated reagent control predicts the stereochemistry of both epoxidations. Molecular sieves maintain a dry active titanium–tartrate system.[ref]
The first practical method for asymmetric epoxidation by Tsutomu Katsuki and K. Barry Sharpless. JACS 102, pp. 5974–5976: tartrate-controlled allylic-alcohol epoxidation
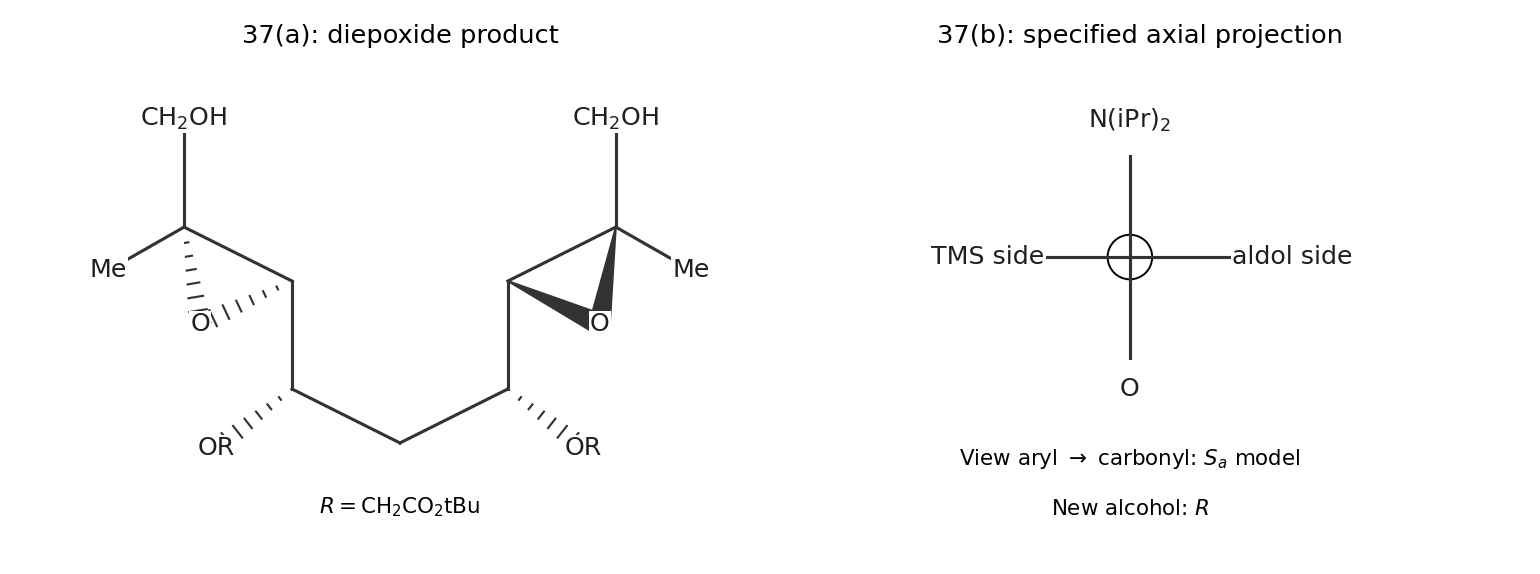
Original facial-selectivity and axial-projection solution models
. Details. 
acetone first forms a proline enamine by reversible iminium formation and alpha-deprotonation. The enamine attacks the aromatic aldehyde in a hydrogen-bonded arrangement involving the catalyst's carboxylic acid. hydrolysis of the resulting iminium releases the beta-hydroxyketone and regenerates proline. The usual L-proline arrangement gives the secondary alcohol.
The sterically crowded aryl–amide axis is selected at the same bond-forming step: rapid rotation in the aldehyde permits the catalyst to react preferentially with the conformer that puts the smaller carbonyl-oxygen side of the amide towards the approaching aldol transition structure and the bulky diisopropylamino side away. A chair-like enamine–aldehyde model minimizes the interaction of the amide with the catalyst-side axial hydrogen; reversing the amide orientation creates the less favourable diastereomeric transition state. Replacing planar CHO by tetrahedral CH(OH)CHCOMe then raises the rotation barrier and traps the axis. This is a dynamic kinetic resolution, not resolution of two configurationally stable starting atropisomers.
To specify the predicted major axis without relying on a flat bond-line formula, look from the aromatic ipso carbon towards the amide carbonyl carbon. Put the higher-priority aryl path, the trimethylsilyl-bearing ortho side, at nine o'clock. The preferred model puts the carbonyl oxygen at approximately six o'clock and the diisopropylamino group at twelve o'clock. This is using the near-to-far high-priority convention; oxygen outranks nitrogen at the far end. Coupled with the alcohol assignment, the model predicts . The drawing defines the configuration even if an alternative viewing direction is used. The dynamic-resolution mechanism is established by the primary aldol study; the product-axis assignment here is a stereochemical-model prediction for the printed substrate.[ref]
Dynamic kinetic resolution of atropisomeric amides by Vincent Chan and others. Organic Letters 6, pp. 2051–2053: simultaneous alcohol and amide-axis control
An unsaturated Rh(I)–BINAP complex undergoes oxidative addition of the aldehydic C–H bond, giving a Rh(III) acyl hydride. Coordination of the tethered alkene brings it into the chiral coordination sphere. hydride migration to the internal, tert-butyl-substituted alkene carbon places Rh on the terminal methylene. The resulting acyl–alkyl complex has the connectivity required for a five-membered ring. reductive elimination between its acyl and alkyl carbons closes the C–C bond, releases the cyclopentanone and regenerates Rh(I).
The aldehyde carbon becomes the ketone carbonyl carbon; its hydrogen ultimately reaches the alkene-derived internal carbon. The original tert-butyl substituent remains attached to the internal alkene carbon, which becomes the beta stereocentre of the cyclopentanone. alkene coordination and insertion favour the chiral pocket arrangement with the bulky tert-butyl group in the less crowded region of the BINAP complex. For the stated (S)-BINAP system the observed product is the beta-tert-butylcyclopentanone, equivalently the product with tert-butyl on the solid-wedge face in the supplied ring orientation. Its priorities are tert-butyl, the shorter carbonyl-side ring path, the other ring path, and hydrogen. The ligand's S label alone is not a general rule that every product stereocentre must be S; the substrate orientation in the catalyst must also be specified. No decarbonylation is needed in the productive hydroacylation cycle.
One valid asymmetric route uses N-Boc-(R)-proline as a chiral starting material. Convert its acid to the Weinreb amide with N,O-dimethylhydroxylamine and a coupling reagent such as EDC, then add phenylmagnesium bromide at low temperature. chelation of the magnesium tetrahedral intermediate prevents a second addition; aqueous workup releases the phenyl ketone. Reduce it with sodium borohydride, separate the resulting alcohol diastereomers, and retain the one with configuration at both pyrrolidine C2 and the benzylic carbinol. This is an asymmetric synthesis through a pre-existing enantiopure centre, even though the final reduction need not be perfectly diastereoselective. If the other protected alcohol is isolated, Mitsunobu esterification with a carboxylic acid followed by ester hydrolysis inverts just the carbinol centre. Keeping nitrogen Boc-protected during that inversion avoids neighbouring-amine participation. Mild Boc removal followed by reductive benzylation with benzaldehyde and sodium triacetoxyborohydride gives D, the amino alcohol. Use conditions that avoid epimerizing the amino ketone.
For ring expansion, activate D's alcohol with methanesulfonyl chloride and a tertiary amine base. Its own tertiary nitrogen attacks the benzylic carbon from the back, displacing mesylate with inversion and making a fused aziridinium ion. Acetate, supplied as a soluble acetate salt, attacks the original pyrrolidine C2 from the opposite face and breaks the old C2–N bond. The new N–benzylic-C bond is retained, incorporating that side-chain carbon into the six-membered ring. Inversion at the two successively attacked carbons gives the cis acetate/phenyl piperidine E in the displayed orientation. Attack at the benzylic carbon instead would simply open the three-membered ion back to a five-membered-ring derivative; specifying the attacked carbon is the crucial regiochemical step.
Articles by others on the same topic
There are currently no matching articles.