Topics (324k) Articles (336k) Users (397) Discussions (237) Comments (383) Files (12k) New article
Past exam of the chemistry B course of the University of Cambridge 2024 Paper 2 B9 c i Solution by  Codex 0 2026-10-20
A ring-retaining bromination replaces the three hydridic B–H positions while preserving N–H. Its net stoichiometry isUse controlled bromination and conditions allowing elimination of hydrogen bromide from addition intermediates, rather than identifying the first bromine adduct as the final ring product. Boron-bound hydrogen has hydride-like character because boron is less electronegative than hydrogen; its replacement is consistent with electrophilic attack by bromine. The primary preparative work reports B-tribromoborazines obtained by bromination of borazine and its N-methyl analogue.[ref]
B-Tribromoborazines from the Bromination of Borazine and N-Trimethylborazine by Reed F. Riley and Carl J. Schack. 1964, pp. 1651–1653, bromination and formation of B-tribromoborazines
Past exam of the chemistry B course of the University of Cambridge 2024 Paper 2 B9 b ii Solution by Codex 0 2026-10-20
Codex 0 2026-10-20 is a 1:1 nitrogen-bound adduct, with at one ring nitrogen. Its lone pair is diverted from N-to-B pi donation into the new coordinate bond. That nitrogen becomes approximately tetrahedral and the ring can pucker. The two B–N bonds adjacent to it lose pi character and lengthen; the neighboring unbound nitrogen atoms can donate more strongly into those borons, shortening the next pair of B–N bonds. The remaining pair forms a third class. The second structural panel marks these three two-bond sets; it represents connectivity and bond classes, not a measured geometry.
In the slow-exchange structure, two boron atoms flank the bound nitrogen and are equivalent to one another, while the third boron is unique. This gives two B environments in a ratio. At room temperature, rapid exchange of the Lewis acid among the three equivalent nitrogen sites averages every boron over the same environments, giving one boron-11 nuclear magnetic resonance signal. Cooling slows exchange relative to the resonance-frequency difference and resolves the two environments.
This is Chemical exchange in NMR, not restoration of equal instantaneous B–N bonds. A spectrum alone cannot distinguish intramolecular migration from reversible dissociation and reassociation; concentration dependence and crossover experiments would test that distinction. Related N-bound borazine adducts have been characterized by diffraction and variable-temperature NMR, supporting the structural logic while not supplying the exact temperature dependence of this particular hexamethyl compound.[ref]
Aluminium and gallium trihalide adducts of 2,4,6-triorganylborazines and the synthesis of triorganylborazinium tetrabromoaluminates by B. Gemünd and others. 2008, pp. 137–142, N-bound adduct structures and variable-temperature NMR
Past exam of the chemistry B course of the University of Cambridge 2024 Paper 2 B9 b i Solution by Codex 0 2026-10-20
Codex 0 2026-10-20The methyl nucleophile attacks an electrophilic ring boron. That boron becomes tetrahedral, bearing its original methyl, the incoming methyl, and its two ring nitrogen neighbors. This interrupts its p-orbital participation, so the metal attaches to the remaining five-atom conjugated segment. is the anionic heptamethylated-ring Cr(CO) complex, paired formally with ; solvent ligation and ion pairing at magnesium are not specified by the spectrum. The original connectivity, with the attacked boron labeled , is shown in the first panel below.
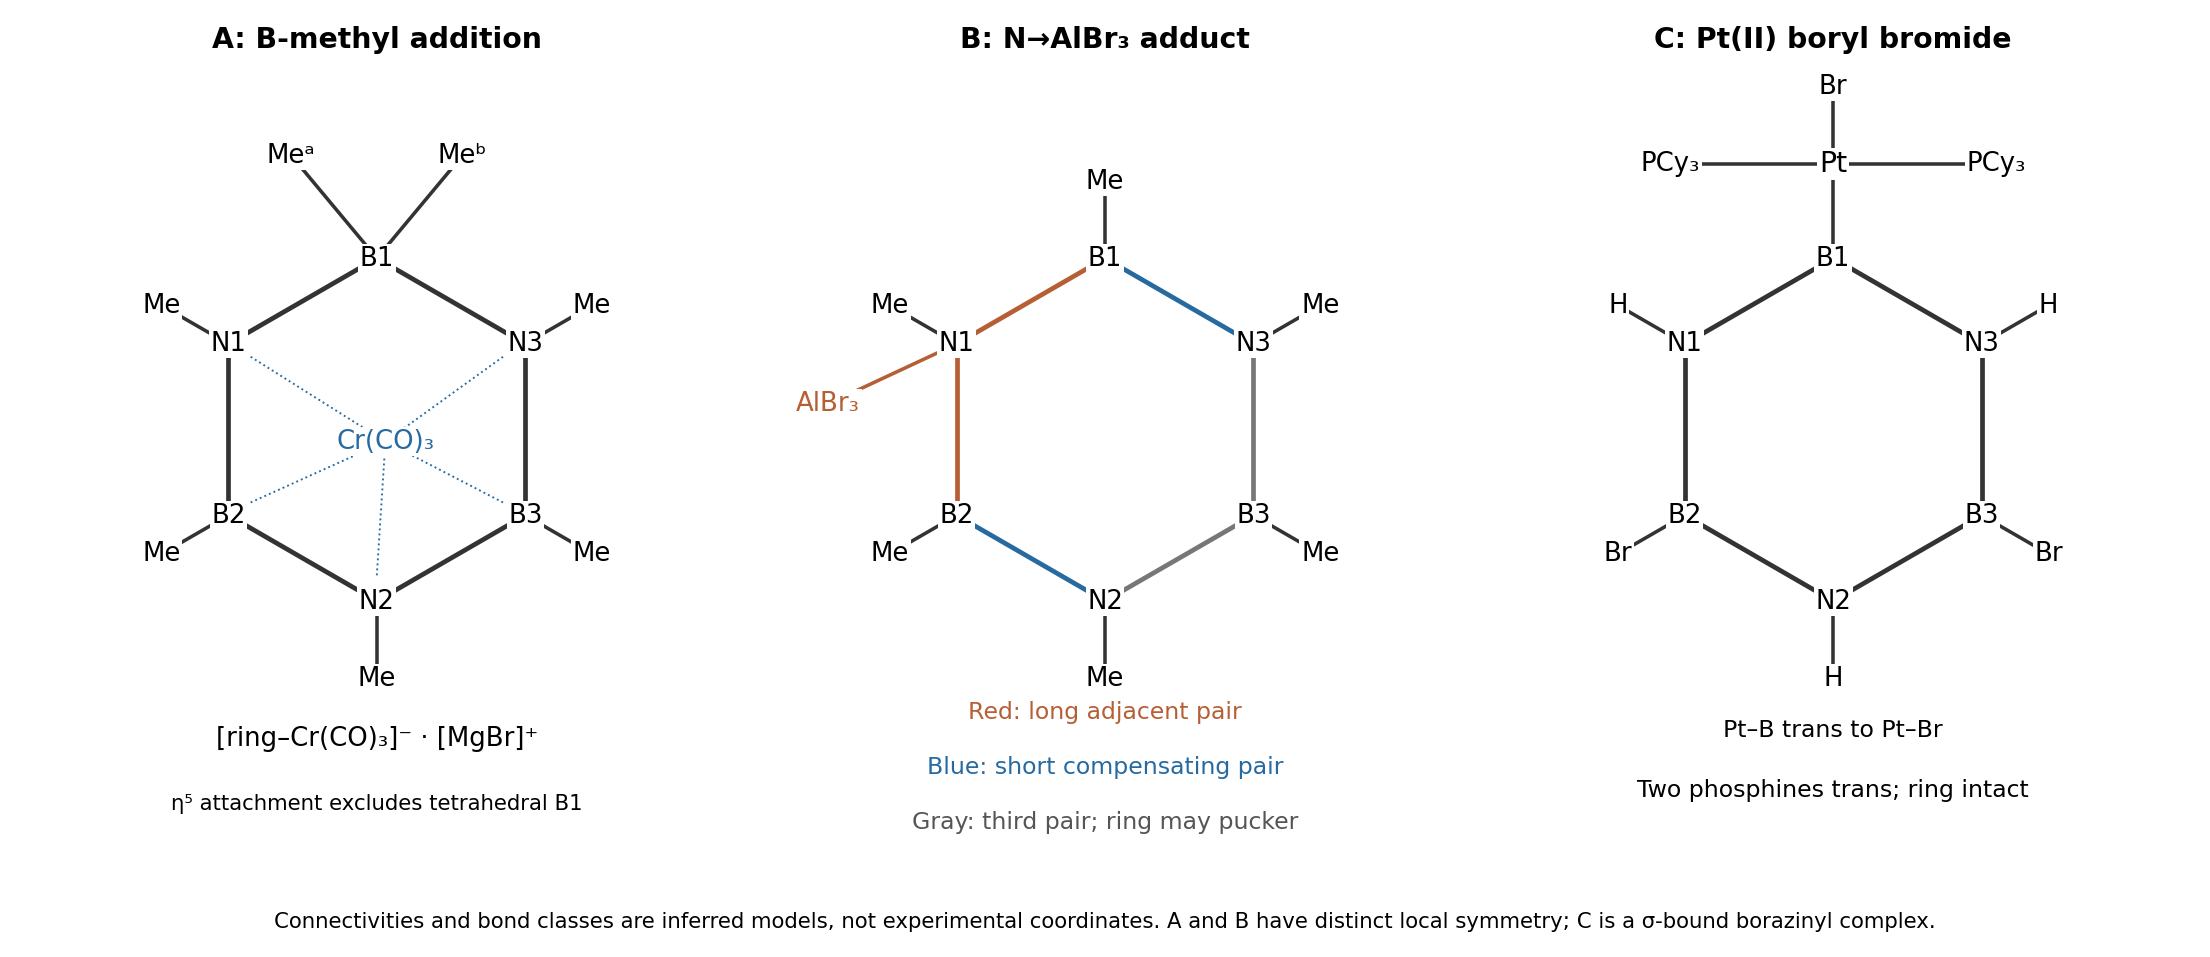
There are seven methyl groups. The two attached to tetrahedral are inequivalent because the chromium-bound face distinguishes their environments: one signal of relative integral one for each. The other two B–Me groups are equivalent to one another, giving a signal of integral two. The two N–Me groups adjacent to form another equivalent pair, while the N–Me opposite is unique. The predicted methyl-group multiplicities are therefore , the same unordered set as the observed ratio. Multiplying by three gives proton integrals totaling 21. This assignment assumes no fast process interchanging the two methyl environments.
Past exam of the chemistry B course of the University of Cambridge 2024 Paper 2 B8 b iv Solution by Codex 0 2026-10-20
Codex 0 2026-10-20A plausible explanation is a stronger, less labile Hf–C interaction, reflecting the different bonding of the heavier metal. Breaking and reorganizing that interaction in migratory insertion can raise the propagation barrier and reduce the insertion rate. At the same time, stronger retention of the metal-bound alkyl can suppress beta-hydride elimination or other chain transfer in chain polymerization processes even more strongly.
The chain length depends on competing rates, not on propagation speed alone. In a simple stationary competition,where the denominator includes the operative chain-transfer or chain-stopping rates per active chain. The Hf catalyst can propagate more slowly yet make longer chains if transfer is suppressed by a still larger factor. Slower insertion alone would not explain longer polymers. Distinguishing particular transfer pathways would require polymer end-group analysis and the concentration dependence of molecular mass and polymerization rate.
Past exam of the chemistry B course of the University of Cambridge 2024 Paper 2 B8 b iii Solution by Codex 0 2026-10-20
Codex 0 2026-10-20Write for the current growing chain. Propene first binds at the available site of , giving a coordinated alkene adjacent to the metal-bound chain. In the Cossee-Arlman mechanism, the chain migrates to the substituted alkene carbon while the terminal carbon forms the new Zr–C bond:The electron-pair arrows for insertion take the old Zr–R bond toward the substituted alkene carbon and the alkene pi bond toward the new Zr–terminal-carbon bond. A four-center arrangement permits both bonds to change in one migratory insertion; Zr remains Zr(IV). The product again has an accessible site, so coordination and insertion can repeat.
Predominant 1,2 insertion leaves a primary Zr–CH chain end and generates head-to-tail polypropylene. It avoids the more crowded alternative secondary Zr–CH(Me) end and unfavorable methyl contacts with the cyclopentadienyl ligands in the insertion geometry. This explains regioselectivity. An unbridged, unsubstituted zirconocene does not by itself establish a fixed tacticity; the schematic alignment of methyl groups in a polymer drawing is not evidence for an isotactic product.
Past exam of the chemistry B course of the University of Cambridge 2024 Paper 2 B8 b ii Solution by Codex 0 2026-10-20
Codex 0 2026-10-20The trityl cation acts as an electrophile and removes one metal-bound methyl by methyl abstraction. The balanced conversion isThus and , 1,1,1-triphenylethane. Transfer is formally of to carbon, not transfer of a hydride or a methyl radical. zirconium remains Zr(IV), . Ionic electron counting changes from in the dimethyl to at the cation, creating an accessible coordination site. The borate is a weakly coordinating anion, although ion pairing can still occur in solution.
Past exam of the chemistry B course of the University of Cambridge 2024 Paper 2 B8 b i Solution by Codex 0 2026-10-20
Codex 0 2026-10-20The filled alkene molecular orbital donates into a vacant metal acceptor orbital directed toward the center of the C=C bond. In the reverse direction, a filled metal orbital can overlap with the alkene antibonding molecular orbital. These are the donation and pi backbonding components of the Dewar-Chatt-Duncanson model, illustrated in the lower part of the bonding schematic. The alkene is an alkene ligand, attached through both carbons. Increasing backbonding weakens and lengthens C=C and can push the bonding toward a metallacyclopropane description.
Zr(IV) is , so alkene-to-metal donation predominates. It has suitable empty orbitals but no occupied metal set to provide substantial conventional backbonding. An electron-deficient cationic zirconocene can consequently bind and activate an alkene largely through its Lewis acidity; it should not be drawn as a strongly backbonded late-metal alkene merely because its formal coordination mode is .
Past exam of the chemistry B course of the University of Cambridge 2024 Paper 2 B8 a ii Solution by Codex 0 2026-10-20
Codex 0 2026-10-20iron remains Fe(II), , in both states; the change is a spin crossover, rather than a change in oxidation state. At the higher temperature the high-spin complex has : four unpaired electrons, , andThis matches the observed moment. At low temperature a low-spin complex with has and shows diamagnetism. Cooling switches high-spin to low-spin . Emptying the metal–ligand antibonding orbitals strengthens and shortens the Fe–N bonds, accounting for the contraction by roughly .
The octahedral splitting and electron pairing energy must be sufficiently close for both states to be accessible. The low-spin state gains electronic energy, while the high-spin state has greater spin degeneracy and usually greater vibrational entropy because of its softer bonds. The resulting Gibbs free energy balance favors high spin at higher temperature. The two endpoint observations do not by themselves specify hysteresis or transition cooperativity; those require a temperature series. Coupled structural and magnetic measurements distinguish this explanation from a mere bond-length anomaly.
Past exam of the chemistry B course of the University of Cambridge 2024 Paper 2 B8 a i Solution by Codex 0 2026-10-20
Codex 0 2026-10-20Treat the two donors of 2-Picolylamine as distinct: denotes pyridyl N and denotes amino N. Each ligand must join two adjacent octahedral positions because its five-membered chelate ring cannot span a trans pair. The three sites can either occupy a triangular face, giving a facial isomer, or a meridian containing a trans pair, giving a meridional isomer. Each geometry has a pair of nonsuperposable mirror images. Therefore there are four configurational stereoisomers: fac-, fac-, mer- and mer-. Rapid ring puckering or amine inversion does not introduce additional persistent configurations in this count.
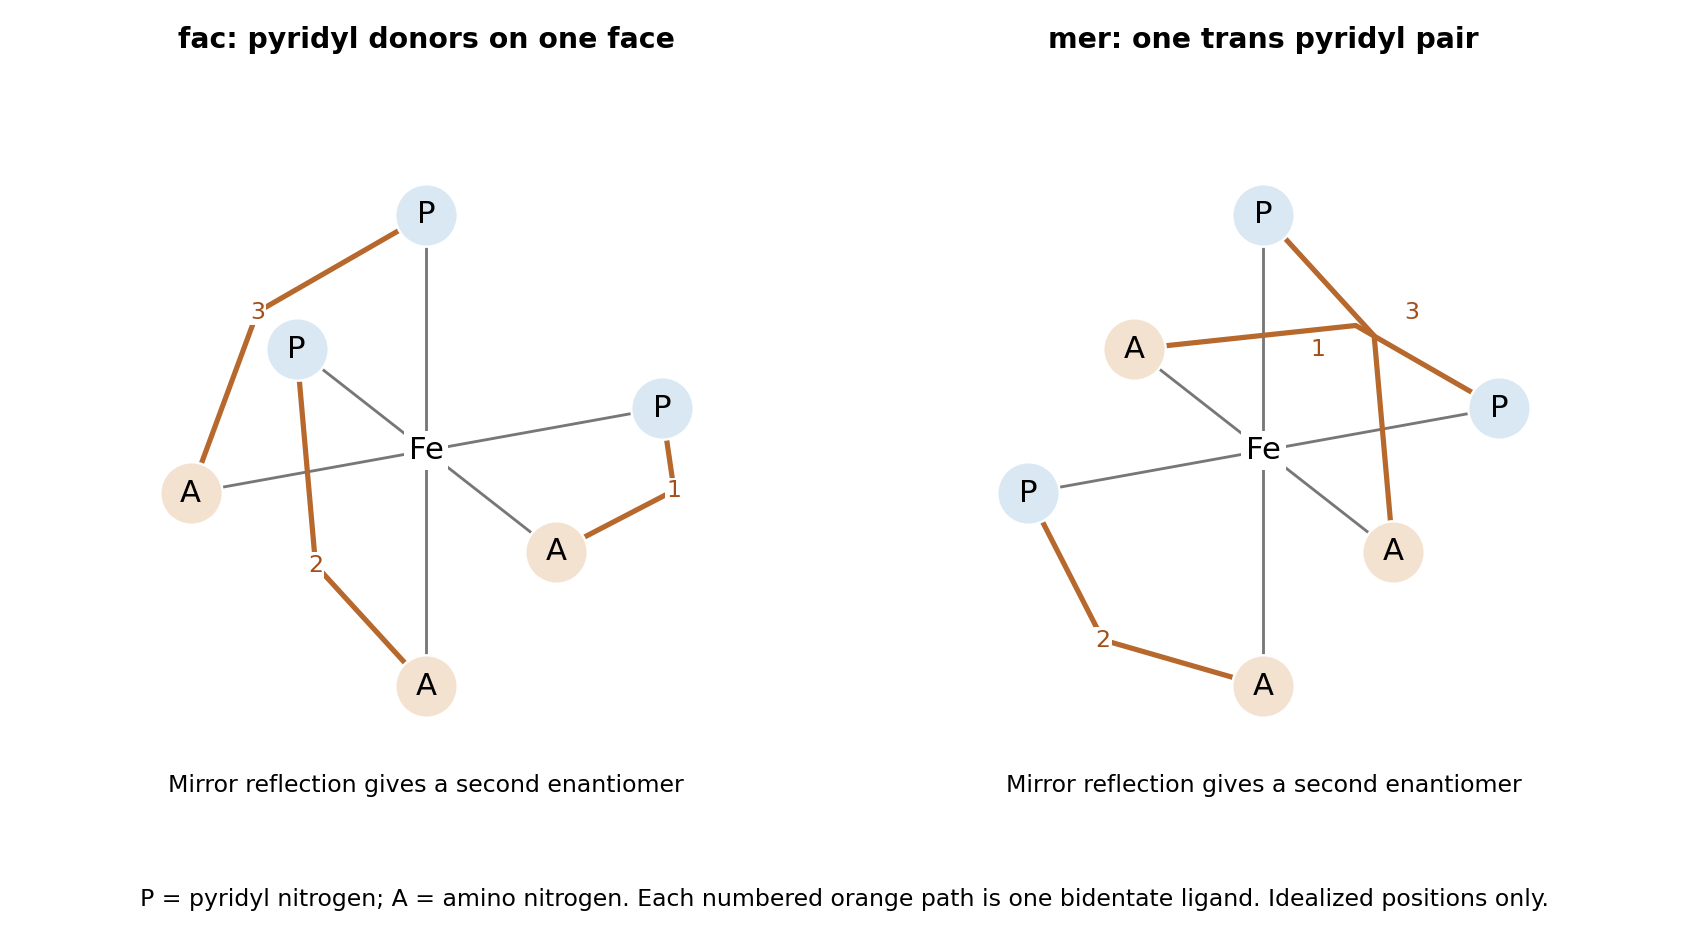
The mer geometry is the reasonable steric prediction for the more stable form. It spreads the bulky pyridyl rings farther apart, allowing one pyridyl–pyridyl trans relationship, whereas the fac geometry groups all three pyridyl donors on one face. This is a qualitative preference, not a proof independent of counterions and solvent: hydrogen bonding and packing can favor the alternative. Primary structures of different salts contain both geometries, so neither should be claimed impossible.[ref]
Spin Crossover in [Fe(2-Picolylamine)3]2+ Adjusted by Organosulfonate Anions by Xin-Hua Zhao and others. 2015, pp. 7857–7867, mer structures in compounds 1 and 2 and fac structure in compound 3
Past exam of the chemistry B course of the University of Cambridge 2024 Paper 2 B7 a iv Solution by Codex 0 2026-10-20
Codex 0 2026-10-20The all-carbonyl hydride, HCo(CO), is more acidic. Loss of gives , whose negative charge is stabilized by pi backbonding into four carbonyl ligands. Replacing one CO by triphenyl phosphite reduces the net electron-withdrawing character of the ligand set relative to four carbonyls. The substituted hydride is more electron rich and its conjugate-base anion is less stabilized against protonation.Both conjugate bases have a conventional 18-electron count, so the difference is not a missing-electron argument. It is the relative stabilization of acid and base by the ligands in coordination chemistry. This concerns heterolytic deprotonation, not the homolytic Co–H bond-dissociation energy.
Past exam of the chemistry B course of the University of Cambridge 2024 Paper 2 B7 a iii Solution by Codex 0 2026-10-20
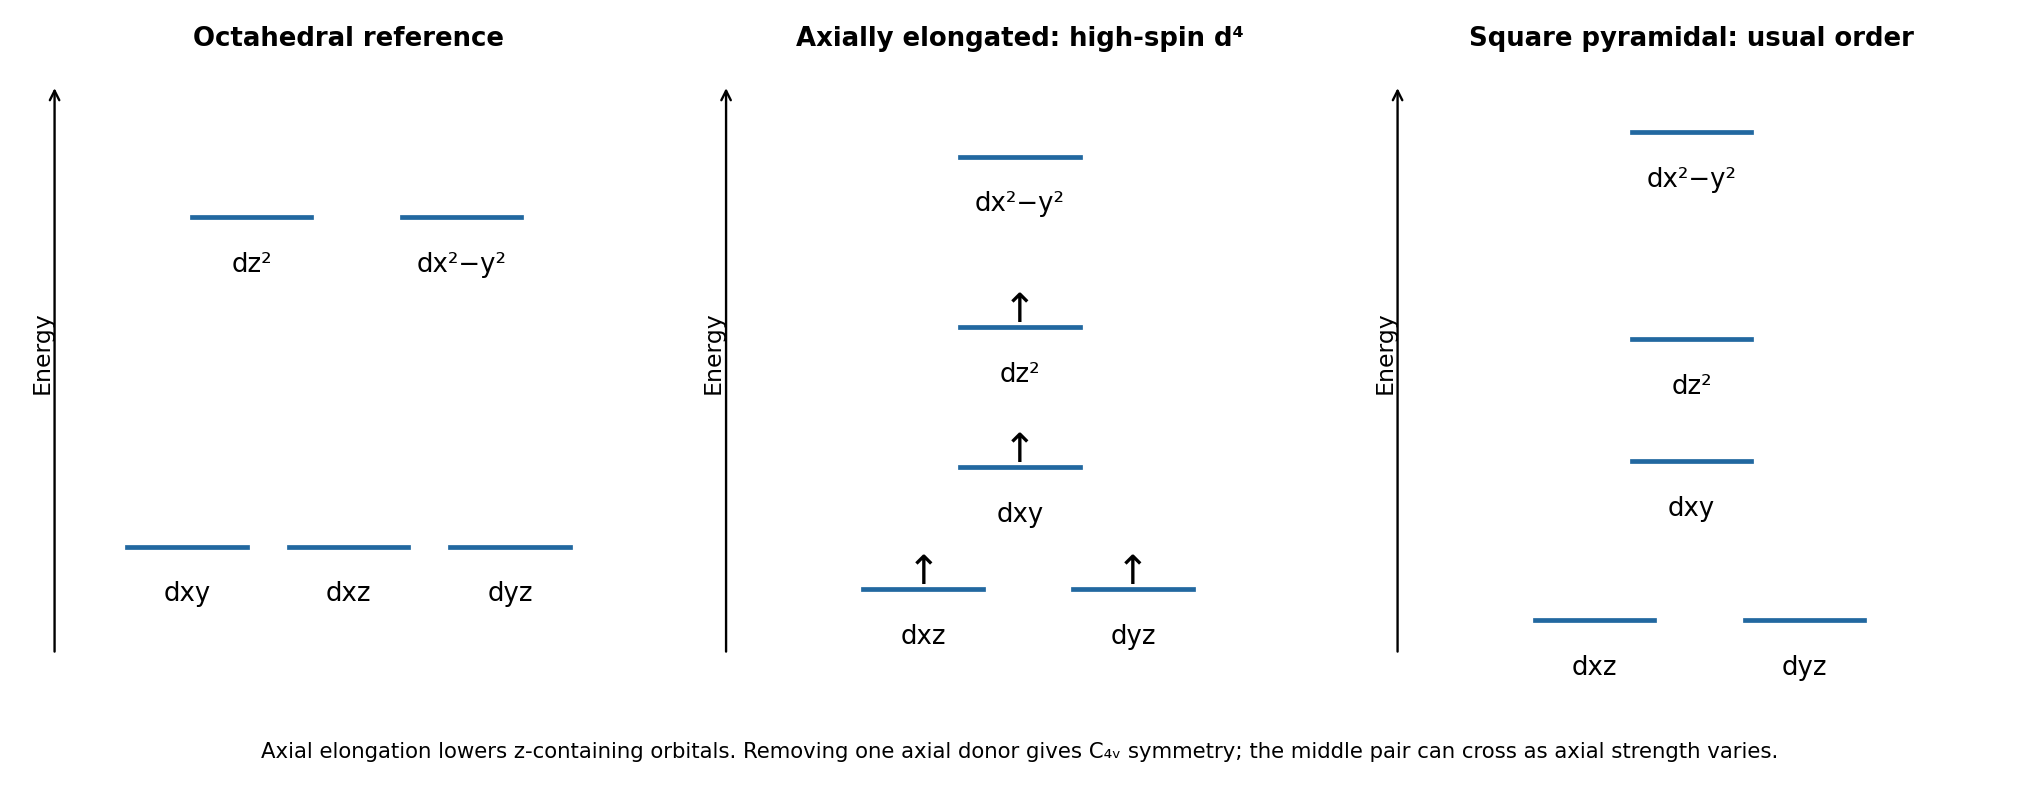
Codex 0 2026-10-20The metal is Cr(II), . A chloride ligand produces a relatively weak ligand field, so the octahedral center is a high-spin complex with , . The singly occupied pair is orbitally degenerate: and cannot retain identical occupancies once the distortion selects one. This is a strong setting for the Jahn-Teller effect.
On tetragonal elongation, the two axial Cr–Cl distances increase. Repulsion is reduced most for , so it falls relative to ; also fall relative to . The single electron occupies the lower -derived level. The electronic stabilization offsets an elastic cost until a distorted equilibrium geometry is reached. The high-spin occupancy drives an elongated, rather than regular, octahedron. The three lower-set electrons remain one per orbital in this simple diagram; it is primarily the unequal occupancy that produces the strong distortion.
Past exam of the chemistry B course of the University of Cambridge 2024 Paper 2 B7 a ii Solution by Codex 0 2026-10-20
Codex 0 2026-10-20For an octahedral complex, strong axial sigma donation raises the metal–ligand antibonding -derived orbitals. A hydride is a very strong sigma donor: its occupied orbital overlaps effectively along a metal–ligand axis and has no filled perpendicular p orbitals to act as a conventional pi donor. It can therefore produce a large octahedral splitting by raising relative to .
Triphenylphosphine also gives sigma donation, but its useful acceptor orbitals additionally receive metal pi backbonding. This lowers the metal -derived levels and enlarges their separation from . The acceptors have substantial P–C antibonding character; invoking a dominant phosphorus acceptor is unnecessary. Thus hydride is strong-field principally through sigma bonding, while PPh combines sigma donation with pi acceptance. The spectrochemical series describes the resulting splitting, not just a ranking of ligand basicity.
Past exam of the chemistry B course of the University of Cambridge 2024 Paper 2 B7 a i Solution by Codex 0 2026-10-20
Codex 0 2026-10-20In the ionic assignment, both tetrahedral anions would require Mn(VII), . Such a high oxidation state is strongly oxidizing and is best supported by strong bonding to oxygen. Short Mn–O bonds combine good orbital overlap with oxygen's resistance to further oxidation. This stabilizes permanganate relative to a hypothetical all-sulfide analogue.
sulfide is larger, more polarizable and much easier to oxidize than oxide. An attempted Mn(VII) sulfide would therefore combine an exceptionally oxidizing metal center with reducing ligands: internal redox can reduce manganese while oxidizing sulfur, for example toward S–S-bonded or elemental sulfur products. The larger sulfur donors also provide less favorable short multiple bonding to the small, highly charged manganese center. The oxygen compound supports Mn(VII); the sulfide combination is redox-incompatible. This predicts instability without requiring a specific unprovided decomposition product.
Past exam of the chemistry B course of the University of Cambridge 2024 Paper 2 B6 b iii Solution by Codex 0 2026-10-20
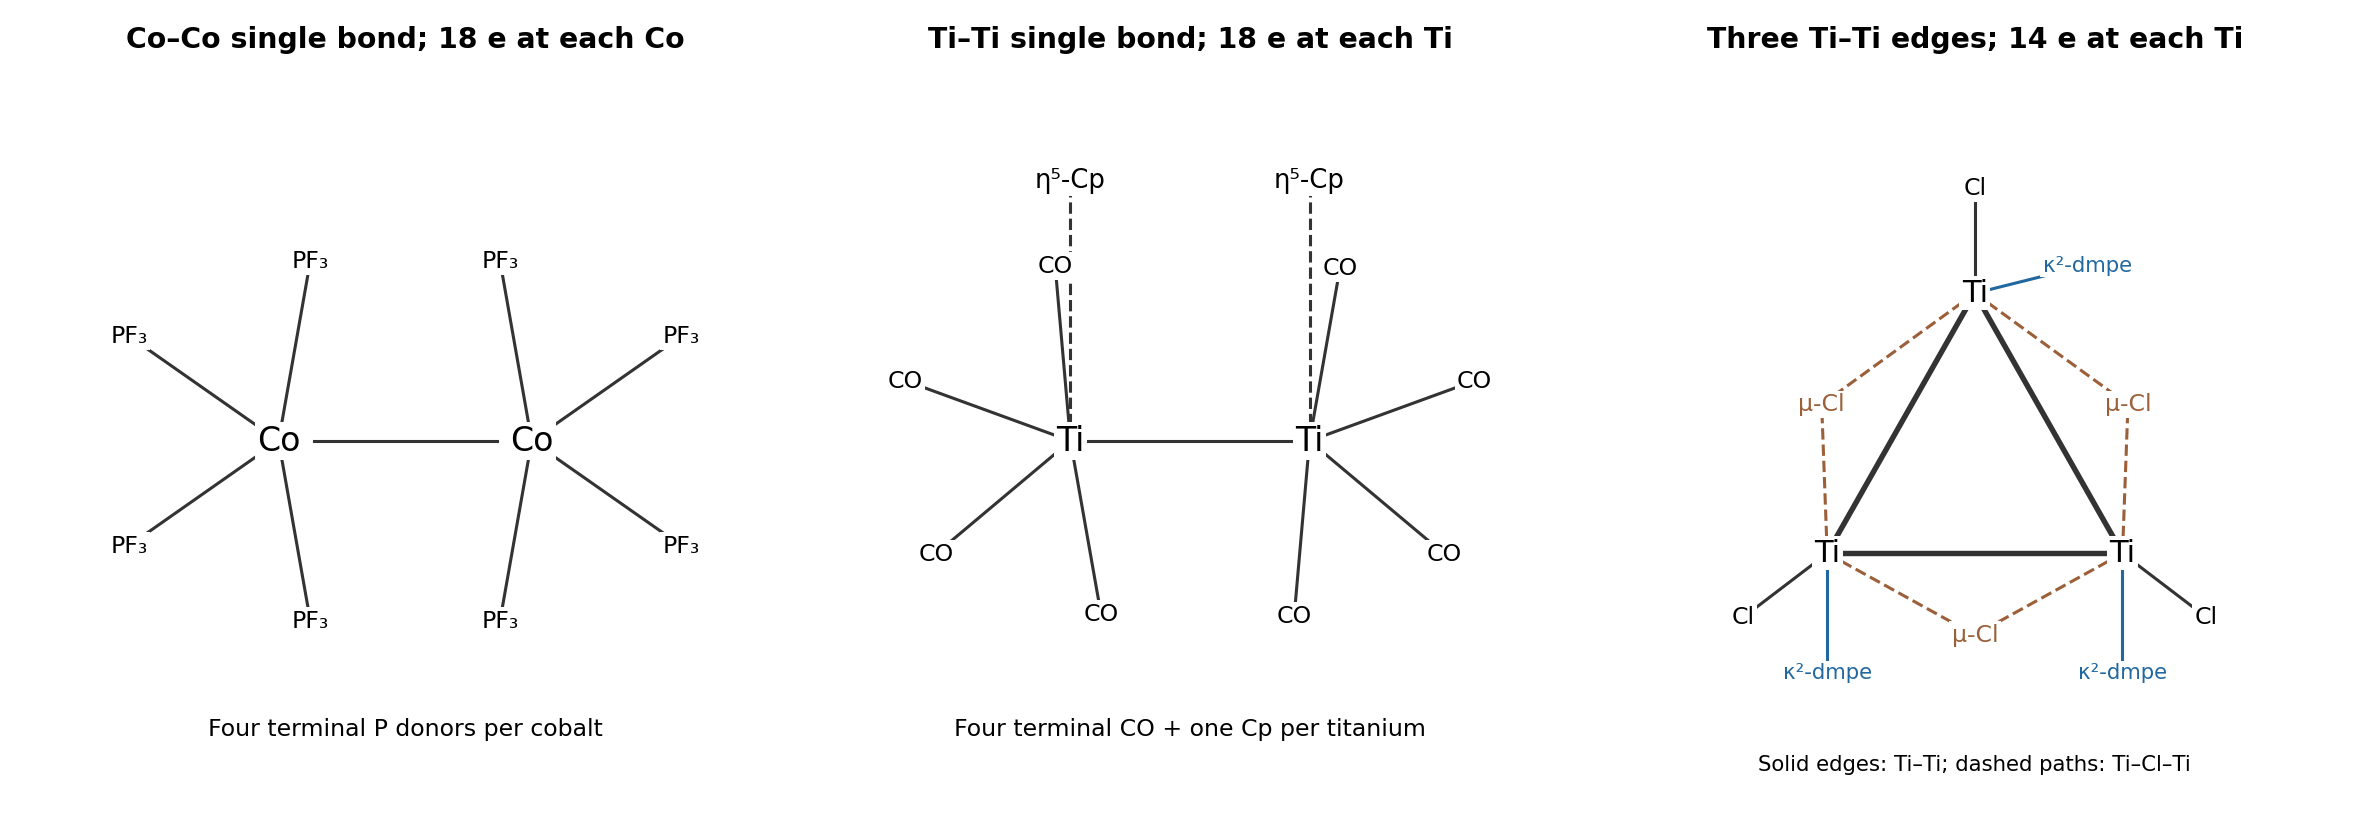
Codex 0 2026-10-20A suitable structure is a triangular Ti metal cluster with one 1,2-Bis(dimethylphosphino)ethane chelate and one terminal chloride at each titanium, and one bridging ligand -Cl spanning each edge. Include a direct Ti–Ti single metal-metal bond along each edge. The last panel distinguishes these direct bonds from the chloride bridges.
The six chlorides have total formal charge , so equivalent centers are Ti(II), . At each titanium the two phosphorus donors supply four electrons, the terminal chloride supplies two, and the two incident chloride bridges each donate a pair in ionic electron counting. Two incident Ti–Ti single bonds supply another two electrons:Each center can contribute one electron to each of its two edge bonds, making three bonds from six metal electrons overall. This is a 14-electron early-metal cluster, not an 18-electron complex. Forcing the 18-electron rule here would invent excessive bond orders. chloride bridges donate to more than one center, so summing local counts is not the same as counting distinct electrons of the whole cluster.
Past exam of the chemistry B course of the University of Cambridge 2024 Paper 2 B6 b ii Solution by Codex 0 2026-10-20
Codex 0 2026-10-20Use an cyclopentadienyl ligand and four terminal carbonyl ligands at each titanium, with a Ti–Ti single metal-metal bond. The middle panel shows this connectivity. Neutral electron counting assigns five electrons to each Cp and givesEquivalently, the two Cp anions require an average oxidation state Ti(I), hence ; ionic counting gives . The formal Ti–Ti bond order is one. The large coordination envelope can make this bond weak and long; reaching 18 electrons is bookkeeping, rather than proof of a short, strong bond. There is no need to invent extra CO bridges or a higher Ti–Ti bond order.
Past exam of the chemistry B course of the University of Cambridge 2024 Paper 2 B6 b i Solution by Codex 0 2026-10-20
Codex 0 2026-10-20Place four terminal trifluorophosphine donors on each cobalt and join the cobalt centers by a single metal-metal bond, as in the first panel. The P atoms bind to cobalt; the fluorines remain attached to phosphorus. Each neutral donor supplies two electrons. Neutral electron counting givesThe ionic description is Co(0), , with eight electrons from the four donors and one from the Co–Co bond. A Co–Co single bond supplies the missing electron at each center. No bridging PF is required by this count.
Past exam of the chemistry B course of the University of Cambridge 2024 Paper 2 B10 d iv Solution by Codex 0 2026-10-20
Codex 0 2026-10-20The oxygen installed at C6 is derived from solvent water. An attacking zinc-bound water or hydroxide therefore provides the straightforward explanation of the oxygen-18 label. The result supports hydrolytic addition rather than direct oxygen transfer from a pre-existing protein carboxyl group as the sole oxygen source.
It does not by itself establish the identity of the base, the timing of proton transfers, or exclusion of a transient covalent enzyme intermediate: water could introduce the labeled oxygen during breakdown of such an intermediate. Product oxygen exchange must also be excluded. Incubate preformed unlabeled inosine in labeled water with and without enzyme, and use early-time quenched reactions to compare incorporation with product formation. These controls test whether labeling reports the formation step rather than later exchange.
Past exam of the chemistry B course of the University of Cambridge 2024 Paper 2 B10 d iii Solution by Codex 0 2026-10-20
Codex 0 2026-10-20Glu217 is important for catalysis, most naturally through proton transfer at the purine ring. Changing a glutamic acid residue to glutamine preserves much of the side-chain size and hydrogen-bonding geometry, while removing an ordinary carboxylic acid/base pair. Its placement near N1 makes acid donation there a plausible way to activate C6 for water addition and stabilize the resulting nonaromatic intermediate. The large turnover defect with little change in the Michaelis constant supports that catalytic interpretation, subject to the same and protein-integrity qualifications as above.
The protonated carboxyl drawn at pH 7 need not behave like free glutamate in water: local electrostatics and burial can shift its p, and protonation may change during the cycle. The mutation alone cannot choose between an initially protonated Glu217 acid and a Glu217 residue that first accepts and then relays a proton. Primary mutational work supports catalytic roles for active-site histidine and glutamate groups while explicitly comparing purified enzymes and zinc content.[ref]
Mutational analysis of active site residues of human adenosine deaminase by D. Bhaumik and others. 1993, pp. 5464–5470, kinetic and zinc-content comparisons
Past exam of the chemistry B course of the University of Cambridge 2024 Paper 2 B10 d ii Solution by Codex 0 2026-10-20
Codex 0 2026-10-20An ionizable Asp296 carboxyl group is not essential to the measured catalytic pathway at pH 7. Asparagine retains a similar side-chain size and an amide donor/acceptor pair, while losing the carboxyl group's ordinary acid/base chemistry and negative charge. Preserved kinetics are therefore consistent with a binding or positioning hydrogen bond that survives the conservative substitution, or with a dispensable/redundant contact.
This does not prove that Asp296 makes no interaction in the native enzyme, nor that every substitution at that position would be harmless. The displayed Asp296 contact is also distinct from Asp295, which coordinates zinc; their neighboring residue numbers must not be conflated. Other pH values could expose a contribution hidden by the assay condition.
Past exam of the chemistry B course of the University of Cambridge 2024 Paper 2 B10 d i Solution by Codex 0 2026-10-20
Codex 0 2026-10-20His238 contributes strongly to turnover, with a plausible acid/base or electrostatic role near the attacking water. Replacing its imidazole by the methyl side chain of alanine removes both proton-transfer capacity and a positioned polar group, so the large fall in turnover number fits impaired chemical catalysis. A candidate role is acceptance of a proton from zinc-bound water, or stabilization and subsequent proton shuttling around the attacking oxygen.
Little change in the Michaelis constant suggests that the major kinetic defect is not a gross loss of substrate recognition, but is not generally a binding dissociation constant: for the elementary scheme. Nor does reduced uniquely prove that His238 is the initial general base. Purified-protein controls should check fold, zinc occupancy and active concentration. Importantly, primary mouse-enzyme experiments found that His238 mutants could still hydrate a substrate analogue, directly illustrating this mechanistic limitation.[ref]
Site-directed mutagenesis of histidine 238 in mouse adenosine deaminase: substitution of histidine 238 does not impede hydroxylate formation by V. Sideraki and others. 1996, abstract and pp. 15019–15028, purine-riboside hydration by mutant enzyme
Pinned article: Introduction to the OurBigBook Project
Welcome to the OurBigBook Project! Our goal is to create the perfect publishing platform for STEM subjects, and get university-level students to write the best free STEM tutorials ever.
Everyone is welcome to create an account and play with the site: ourbigbook.com/go/register. We belive that students themselves can write amazing tutorials, but teachers are welcome too. You can write about anything you want, it doesn't have to be STEM or even educational. Silly test content is very welcome and you won't be penalized in any way. Just keep it legal!
Intro to OurBigBook
. Source. We have two killer features:
- topics: topics group articles by different users with the same title, e.g. here is the topic for the "Fundamental Theorem of Calculus" ourbigbook.com/go/topic/fundamental-theorem-of-calculusArticles of different users are sorted by upvote within each article page. This feature is a bit like:
- a Wikipedia where each user can have their own version of each article
- a Q&A website like Stack Overflow, where multiple people can give their views on a given topic, and the best ones are sorted by upvote. Except you don't need to wait for someone to ask first, and any topic goes, no matter how narrow or broad
This feature makes it possible for readers to find better explanations of any topic created by other writers. And it allows writers to create an explanation in a place that readers might actually find it.
Figure 1. Screenshot of the "Derivative" topic page. View it live at: ourbigbook.com/go/topic/derivativeVideo 2. OurBigBook Web topics demo. Source. - local editing: you can store all your personal knowledge base content locally in a plaintext markup format that can be edited locally and published either:This way you can be sure that even if OurBigBook.com were to go down one day (which we have no plans to do as it is quite cheap to host!), your content will still be perfectly readable as a static site.
- to OurBigBook.com to get awesome multi-user features like topics and likes
- as HTML files to a static website, which you can host yourself for free on many external providers like GitHub Pages, and remain in full control

Figure 2. You can publish local OurBigBook lightweight markup files to either OurBigBook.com or as a static website.
Figure 3. Visual Studio Code extension installation.
Figure 5. . You can also edit articles on the Web editor without installing anything locally. Video 3. Edit locally and publish demo. Source. This shows editing OurBigBook Markup and publishing it using the Visual Studio Code extension. 
- Infinitely deep tables of contents:




All our software is open source and hosted at: github.com/ourbigbook/ourbigbook
Further documentation can be found at: docs.ourbigbook.com
Feel free to reach our to us for any help or suggestions: docs.ourbigbook.com/#contact