For the ideal tetracyanonickelate(II) geometries, the square planar molecular geometry has , the D4h molecular point group, while the tetrahedral molecular geometry has , the Td molecular point group. Collinearity of each Ni–C–N unit means that including the terminal nitrogen atoms does not lower either point group.
For the square, take the Ni–C directions as and perpendicular to the plane. For the tetrahedron, use the conventional cubic axes with ligand directions proportional to . These choices fix the irreducible representation labels used below.
The angular factors of the nickel atomic orbitals transform like quadratic Cartesian functions. In the D4h molecular point group, the assignments areAll five are even under inversion symmetry, hence the suffixes. The pair is mixed by a quarter-turn and spans a two-dimensional irreducible representation.
In the Td molecular point group, the corresponding result isThere is no inversion label because has no inversion centre. In both cases the dimensions add to five.
Choose the phase of each inward-pointing ligand sigma bond donor lobe consistently. An operation taking one donor to another then acts as a permutation on the four donor functions. For the square, label these functions cyclically along . Normalized symmetry-adapted linear combinations areThus the ligand group representation is . Its irreducible representations allow the first two combinations to mix with the matching nickel atomic orbitals; instead matches nickel .
For the tetrahedron in the vertex order specified above, takeThe three latter functions transform like . Hence . These are four ligand symmetry-adapted linear combinations, rather than the five metal functions of part (ii).
Each cyanide contributes charge , so the formal nickel oxidation state is and the metal has . The requested diagram retains only the five principally metal- molecular orbitals; the mainly ligand bonding molecular orbitals lie below them and are omitted.
In the square, points directly towards all four ligand donors and gives the highest, strongly antibonding molecular orbital, . The function also has an allowed interaction, but its in-plane overlap is smaller. Including the allowed nickel mixing gives a principally level with relatively weak antibonding character. The and metal functions have no ligand partner in the sigma bond donor basis and are nonbonding in this minimal model. The useful qualitative ordering is thereforeThe three nonbonding functions are shown together because the sigma-only model does not determine their finer splitting. Additional electrostatic and pi backbonding interactions can change the ordering of the lower levels; molecular symmetry alone does not fix these energy gaps.[ref]
Bonding in Molecules by John E. McGrady. 2022 lecture notes, square-planar sigma-only molecular-orbital discussion, printed pp. 60–61
In the tetrahedron, the ligand combinations interact with the metal functions, producing higher -derived antibonding molecular orbitals. The metal pair has no sigma donor partner and remains nonbonding:The diagram fills both arrangements with eight metal electrons; the square is shown in its fully paired state.
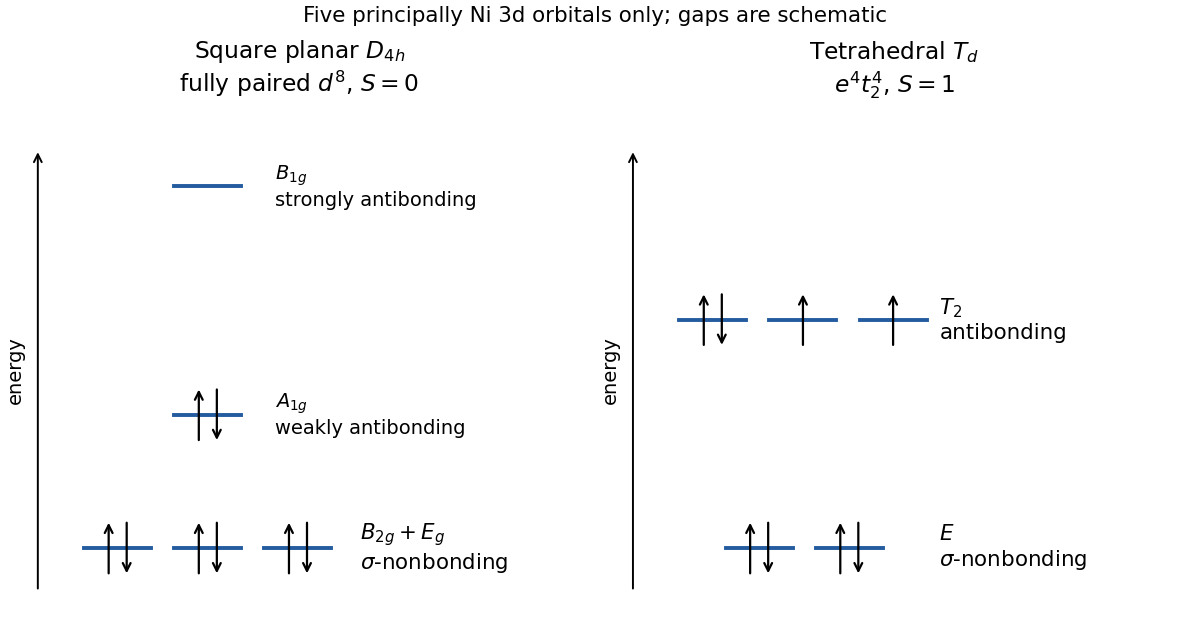
Original qualitative d-derived orbital diagrams for square-planar and tetrahedral nickel(II), with eight electrons and schematic energy gaps
. Details. 
For unpaired electrons, the spin-only magnetic moment iswhere is the Bohr magneton and the second expression uses .
For the square, a large separation of the highest level favours pairing in the lower four molecular orbitals: , , . If the cost of occupying were smaller than the relevant electron pairing energy, a high-spin complex with two unpaired electrons and could instead occur, giving .
For ideal tetrahedral , the occupation is . Hund's rules distribute four electrons over the three degenerate molecular orbitals as one pair and two single occupations. Thus and . Merely increasing the – splitting does not create a conventional paired tetrahedral alternative: four electrons must still occupy .
The D4h molecular point group contains an inversion centre, so infrared and Raman mutual exclusion applies. A normal mode of parity can have vibrational infrared activity, whereas the quadratic polarizability tensor components responsible for Raman vibrational activity have parity. No individual fundamental can be active in both spectra.
The Td molecular point group lacks an inversion centre. In particular, normal modes transform both as electric-dipole components and as off-diagonal polarizability tensor components, so they occur in both spectra. Coincident IR and Raman fundamentals therefore favour the tetrahedral structure; symmetry requires no such coincidences for the square. Weak or unresolved features can conceal coincidences experimentally, while accidental equality of different frequencies is not a violation of mutual exclusion.
There are four internal C–N stretching coordinates. A stretch length is unchanged in sign when a molecular symmetry operation carries a bond to an equivalent bond, so these coordinates have the same permutation group representation as the four ligand sigma functions.
For the square,Only has vibrational infrared activity. The and stretches have Raman vibrational activity. Thus the ideal square gives one IR feature and two Raman features, with no common fundamental. The two components of have one frequency, not two different lines.
For the tetrahedron,The stretch is IR active, while both and are Raman active. Thus the tetrahedron also gives one IR feature and two Raman features, but one feature, , is shared between the spectra. Its three components likewise describe three independent displacements at one frequency.
The separate line counts alone do not distinguish the geometries. Comparing their frequencies does: the tetrahedral IR C–N feature coincides with one Raman C–N feature, whereas the square has no symmetry-enforced coincidence. This assumes resolved fundamentals and excludes accidental equality of unrelated frequencies.
The nine-atom nonlinear ion has vibrational degrees of freedom. To construct its displacement group representation, only atoms left in place by an operation contribute to the character. Each such atom contributes the trace of the operation on :For example, reflection in the square's molecular plane fixes all nine atoms and has trace , giving character ; inversion fixes nickel alone and has trace , giving . Molecular character reduction usesFor the D4h molecular point group, with primed twofold axes along the Ni–C directions, the class order and displacement characters areReducing and removing translations and rotations givesThe vibrational infrared activity belongs to , giving six features. The Raman vibrational activity belongs to , giving seven features. The remaining modes are silent. There are zero shared fundamentals, by infrared and Raman mutual exclusion. Counting dimensions gives IR-active, Raman-active and silent vibrational coordinates, totaling .
For the Td molecular point group, the corresponding calculation starts fromand gives . Subtracting translations and rotations leavesThe fundamentals give four IR features. The fundamentals give eight Raman features, of which four coincide with the IR features. The fundamental is silent. The dimensions are .
Thus the predicted IR/Raman/coincidence counts are for the square and for the tetrahedron. These are numbers of distinct ideal fundamental frequencies: each or multiplet contributes one feature, and different copies of the same irreducible representation generally have different frequencies. Accidental degeneracy, overtones and combination bands are outside this count.
Articles by others on the same topic
There are currently no matching articles.