In atomic units, , and the hydrogen atom hasThe allowed quantum numbers areHere the fixed electron spin quantum number is . The energy eigenvalues are in hartrees; restoring units gives and for the orbital operators and the corresponding factors for spin. The spin-independent hydrogen Hamiltonian does not split the two spin projections.
Keep one normalized spin function fixed and write and , with . Overall constants and the spin factor pass unchanged through the spatial operators.
First, in atomic units, , soFor the squared orbital angular momentum, andTherefore the two terms of the angular operator giveusing . Thus .
Finally, the spatial hydrogen Hamiltonian isFor ,Substitute the already established angular eigenvalue :Hence hartree, consistently with . The angular calculation must precede the radial substitution if one wants to avoid differentiating the entire product repeatedly.
Denote the two complex spatial atomic orbitals by and , retaining the same spin state. With the phases specified by the supplied spherical harmonics, chooseThe two original atomic orbitals are orthonormal, so the squared coefficients add to one and these combinations are normalized and mutually orthogonal. Explicitly,which are real spatial wavefunctions. An overall sign on either orbital has no physical consequence.
These are the familiar and p orbitals. Each has two lobes of opposite wavefunction sign, with an atomic-orbital nodal plane through the atomic nucleus: for and for . The sign is a phase convention, not a positive or negative electric charge; the probability density is nonnegative on both lobes.
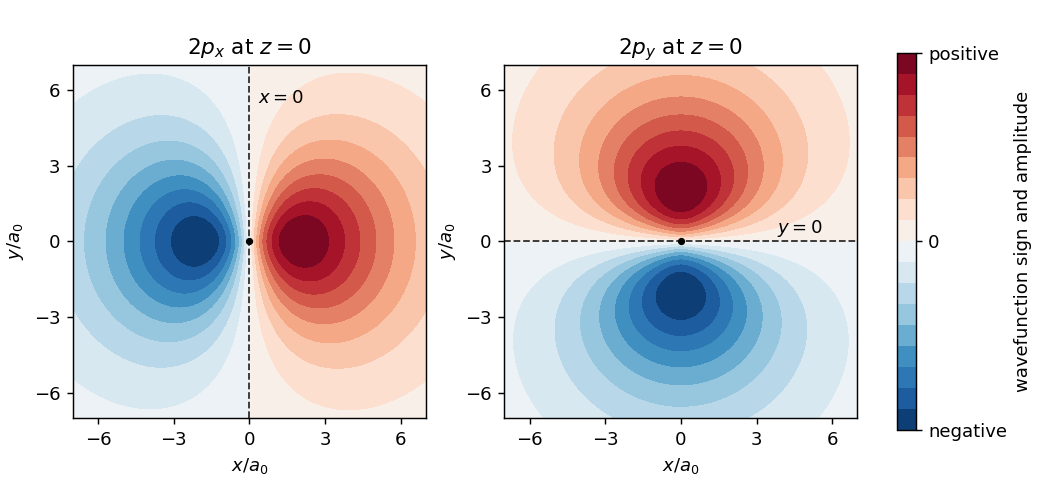
Original sections through the real hydrogen 2p atomic orbitals, showing opposite phases and their nodal planes
. Details. 
Linearity and the shared angular eigenvalue giveUsing the relative phases in part (c),Thus each real orbital remains an energy eigenstate with hartree and has definite orbital angular momentum magnitude , but neither is an eigenfunction of . A measurement of gives or , each with probability . Its mean is zero and its variance is ; a zero mean does not mean that a measurement gives zero.
Let be a complete orthonormal eigenbasis of the Hamiltonian, with and . Expand a nonzero admissible trial wavefunction as . ThenConsequentlyEvery numerator term is nonnegative and the denominator is positive. Equality requires the trial to lie wholly in the ground-state subspace; it need not be an individually chosen eigenvector when the ground energy is degenerate. This is the quantum variational principle, and wavefunction normalization before calculation is optional because the quotient removes the overall scale.
Wavefunction normalization requires . For the normalized probability weight proportional to , the Gaussian integral and its derivatives giveSince , the kinetic energy and potential energy expectations areThusThe second derivative is positive, and the energy diverges at both ends of , so this is the global minimum within the Gaussian function family. The best Gaussian function upper bound isThis is an upper bound, not the exact quartic ground energy.
A gives the same optimum: replacing by merely reparameterizes the same set of positive-width Gaussian functions, so its best bound is unchanged.
B is worse as a ground-energy estimate: it has odd parity and a node at the origin, whereas the ground state of this even confining one-dimensional potential is even and nodeless. It is therefore orthogonal to the exact ground state and belongs to the odd sector, whose lowest energy is already above the ground energy; the extra factor cannot be removed by varying its width. A separate numerical minimization gives a best value about for B, well above the original Gaussian function bound.
C can improve the bound: setting recovers every original Gaussian function, while nonzero adds shape freedom. Minimizing over this larger family can never give a worse optimum, and the additional freedom does lower the quartic-oscillator estimate. Inclusion of a family alone proves “no worse”; improvement is not guaranteed for every conceivable potential, for example one already solved exactly by a Gaussian function.
For any , the Gaussian function is the exact nodeless ground wavefunction of a quantum harmonic oscillator withDirect substitution cancels the terms. An additive constant in the potential would simply shift the eigenvalue.
A natural comparison uses the same optimized width as part (i), . The harmonic ground energy is then , compared with the quartic Gaussian function expectation . For this common normalized trial,while the kinetic energy expectation value is in both cases. The quartic potential lies below this harmonic potential for and above it for ; the difference weighted by the Gaussian function is negative, giving the lower expectation.
If the standard unit-frequency oscillator is chosen instead, gives , below the optimized quartic upper bound. These are different harmonic comparison potentials, not contradictory answers. Crossing potential curves cannot be ordered globally; the comparison must specify the frequency or Gaussian function width used.
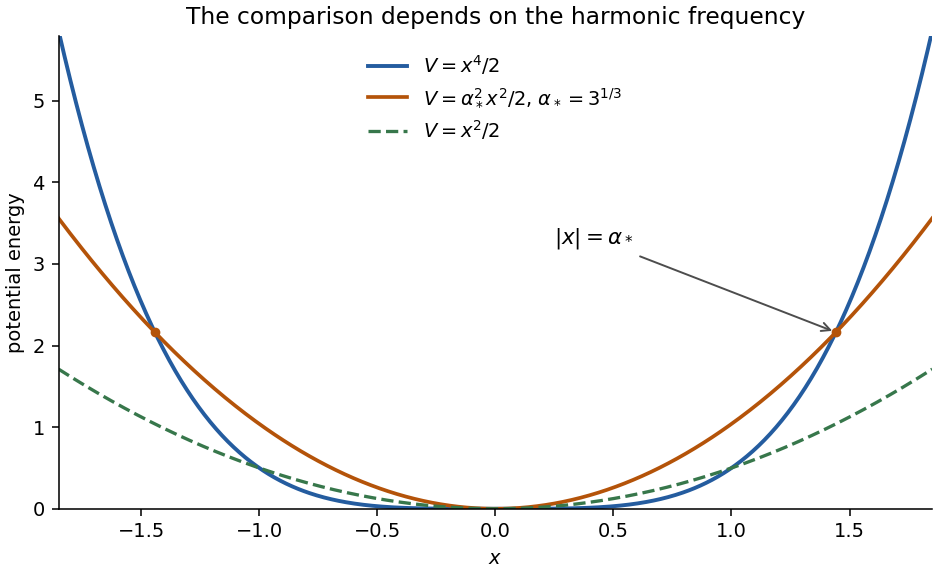
Original comparison of the quartic potential with harmonic potentials at unit frequency and at the optimized Gaussian width
. Details. 
The gross requirement is an anisotropic polarizability tensor, so that changing molecular orientation changes the induced response. A permanent electric dipole moment is not required; a homonuclear diatomic can therefore have a rotational Raman scattering spectrum.
For the inelastic lines of a linear rigid rotor, the detailed rule is , with . The contribution is unshifted Rayleigh scattering, not a rotational Stokes Raman scattering line. Stokes Raman scattering has because the molecule gains rotational energy.
For a transition starting at and ending at , the molecule gainsEnergy conservation therefore gives the scattered-light wavenumbersThe first shifts below the laser are . Adjacent Stokes lines are separated by , while the first line is from the Rayleigh line. The figure uses positive Raman shift ; increasing shift means decreasing scattered-light wavenumber.
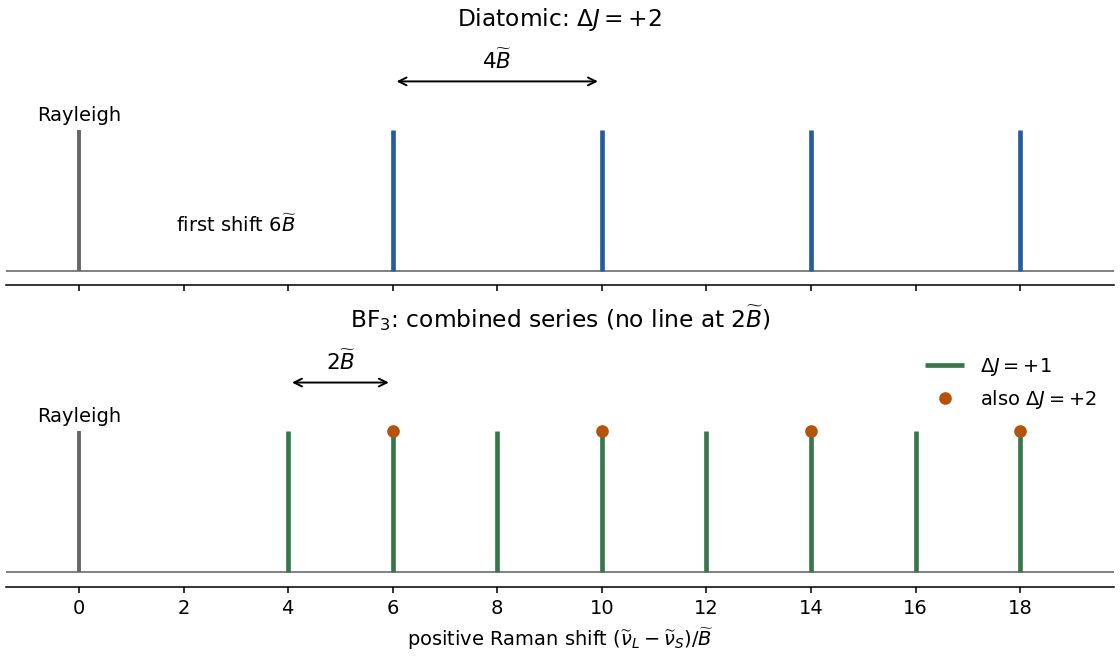
Original line-position sketches for the diatomic and BF3 rotational Raman series; stick heights carry no intensity information
. Details. 
All three fluorine atoms are at perpendicular distance from the axis normal to the molecular plane. Boron lies on the axis, so it contributes zero. ThusThe boron mass does not enter this moment, rather than needing to be approximated or neglected.
In the printed geometry, one B–F bond lies along the in-plane axis and contributes zero. The other two fluorines have perpendicular distance ; boron is again on the axis. ThereforeThe other in-plane principal moment is the same by threefold symmetry. Hence Boron trifluoride is an oblate symmetric top with and .
Because , the terms cancel between the initial and final states. The two positive-shift branches are consequentlyThe branch gives shifts , as for the diatomic. The branch fills in a series spaced by , so the combined nonzero series is .
There is a boundary qualification to the abbreviated rules printed in the question: for an ordinary symmetric polarizability tensor, has zero strength when . Thus does not produce a line; the first transition can start at and has shift . This follows from the rank-two angular-momentum coupling: the all-zero-projection coupling for and vanishes because is odd. If one applies only the printed inequalities without this matrix-element restriction, is a formal position, but its intensity is zero.
The lower panel of the figure in part (a)(ii) marks the two branches and their overlaps. The useful spacing is , not the diatomic value .
The combined boron trifluoride series givesUsing with the wavenumber in inverse metres,With integer fluorine mass and ,Using for the observed spacing would incorrectly treat BF as a linear diatomic rotor and overestimate the bond length by .
For the ideal tetracyanonickelate(II) geometries, the square planar molecular geometry has , the D4h molecular point group, while the tetrahedral molecular geometry has , the Td molecular point group. Collinearity of each Ni–C–N unit means that including the terminal nitrogen atoms does not lower either point group.
For the square, take the Ni–C directions as and perpendicular to the plane. For the tetrahedron, use the conventional cubic axes with ligand directions proportional to . These choices fix the irreducible representation labels used below.
The angular factors of the nickel atomic orbitals transform like quadratic Cartesian functions. In the D4h molecular point group, the assignments areAll five are even under inversion symmetry, hence the suffixes. The pair is mixed by a quarter-turn and spans a two-dimensional irreducible representation.
In the Td molecular point group, the corresponding result isThere is no inversion label because has no inversion centre. In both cases the dimensions add to five.
Choose the phase of each inward-pointing ligand sigma bond donor lobe consistently. An operation taking one donor to another then acts as a permutation on the four donor functions. For the square, label these functions cyclically along . Normalized symmetry-adapted linear combinations areThus the ligand group representation is . Its irreducible representations allow the first two combinations to mix with the matching nickel atomic orbitals; instead matches nickel .
For the tetrahedron in the vertex order specified above, takeThe three latter functions transform like . Hence . These are four ligand symmetry-adapted linear combinations, rather than the five metal functions of part (ii).
Each cyanide contributes charge , so the formal nickel oxidation state is and the metal has . The requested diagram retains only the five principally metal- molecular orbitals; the mainly ligand bonding molecular orbitals lie below them and are omitted.
In the square, points directly towards all four ligand donors and gives the highest, strongly antibonding molecular orbital, . The function also has an allowed interaction, but its in-plane overlap is smaller. Including the allowed nickel mixing gives a principally level with relatively weak antibonding character. The and metal functions have no ligand partner in the sigma bond donor basis and are nonbonding in this minimal model. The useful qualitative ordering is thereforeThe three nonbonding functions are shown together because the sigma-only model does not determine their finer splitting. Additional electrostatic and pi backbonding interactions can change the ordering of the lower levels; molecular symmetry alone does not fix these energy gaps.[ref]
Bonding in Molecules by John E. McGrady. 2022 lecture notes, square-planar sigma-only molecular-orbital discussion, printed pp. 60–61
In the tetrahedron, the ligand combinations interact with the metal functions, producing higher -derived antibonding molecular orbitals. The metal pair has no sigma donor partner and remains nonbonding:The diagram fills both arrangements with eight metal electrons; the square is shown in its fully paired state.
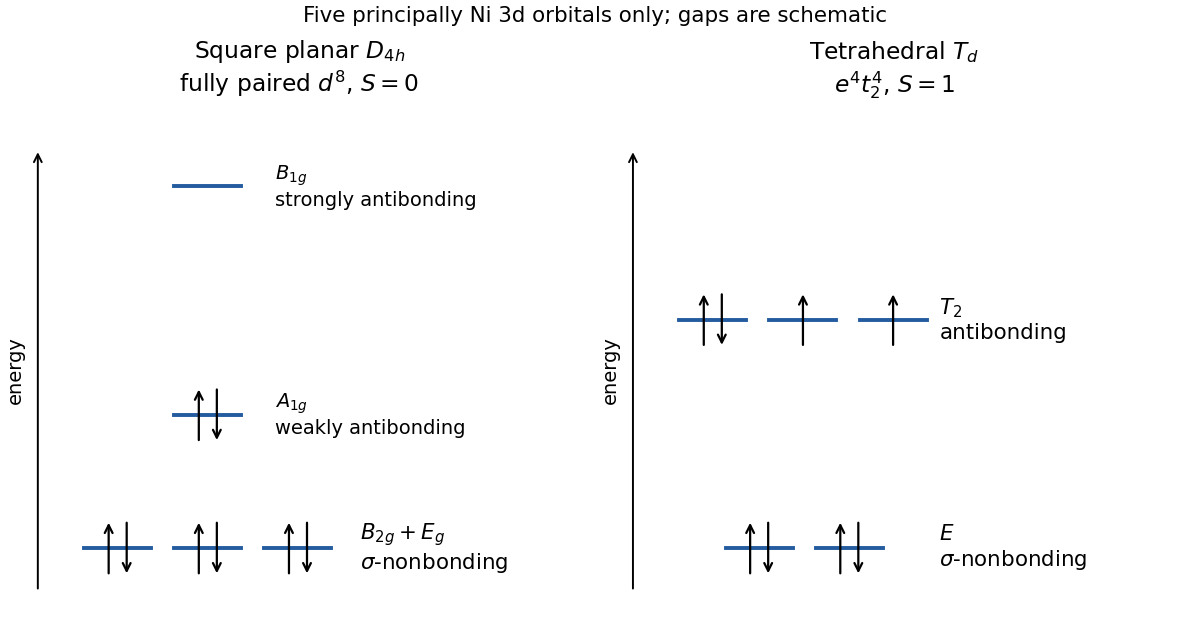
Original qualitative d-derived orbital diagrams for square-planar and tetrahedral nickel(II), with eight electrons and schematic energy gaps
. Details. 
For unpaired electrons, the spin-only magnetic moment iswhere is the Bohr magneton and the second expression uses .
For the square, a large separation of the highest level favours pairing in the lower four molecular orbitals: , , . If the cost of occupying were smaller than the relevant electron pairing energy, a high-spin complex with two unpaired electrons and could instead occur, giving .
For ideal tetrahedral , the occupation is . Hund's rules distribute four electrons over the three degenerate molecular orbitals as one pair and two single occupations. Thus and . Merely increasing the – splitting does not create a conventional paired tetrahedral alternative: four electrons must still occupy .
The D4h molecular point group contains an inversion centre, so infrared and Raman mutual exclusion applies. A normal mode of parity can have vibrational infrared activity, whereas the quadratic polarizability tensor components responsible for Raman vibrational activity have parity. No individual fundamental can be active in both spectra.
The Td molecular point group lacks an inversion centre. In particular, normal modes transform both as electric-dipole components and as off-diagonal polarizability tensor components, so they occur in both spectra. Coincident IR and Raman fundamentals therefore favour the tetrahedral structure; symmetry requires no such coincidences for the square. Weak or unresolved features can conceal coincidences experimentally, while accidental equality of different frequencies is not a violation of mutual exclusion.
There are four internal C–N stretching coordinates. A stretch length is unchanged in sign when a molecular symmetry operation carries a bond to an equivalent bond, so these coordinates have the same permutation group representation as the four ligand sigma functions.
For the square,Only has vibrational infrared activity. The and stretches have Raman vibrational activity. Thus the ideal square gives one IR feature and two Raman features, with no common fundamental. The two components of have one frequency, not two different lines.
For the tetrahedron,The stretch is IR active, while both and are Raman active. Thus the tetrahedron also gives one IR feature and two Raman features, but one feature, , is shared between the spectra. Its three components likewise describe three independent displacements at one frequency.
The separate line counts alone do not distinguish the geometries. Comparing their frequencies does: the tetrahedral IR C–N feature coincides with one Raman C–N feature, whereas the square has no symmetry-enforced coincidence. This assumes resolved fundamentals and excludes accidental equality of unrelated frequencies.
The nine-atom nonlinear ion has vibrational degrees of freedom. To construct its displacement group representation, only atoms left in place by an operation contribute to the character. Each such atom contributes the trace of the operation on :For example, reflection in the square's molecular plane fixes all nine atoms and has trace , giving character ; inversion fixes nickel alone and has trace , giving . Molecular character reduction usesFor the D4h molecular point group, with primed twofold axes along the Ni–C directions, the class order and displacement characters areReducing and removing translations and rotations givesThe vibrational infrared activity belongs to , giving six features. The Raman vibrational activity belongs to , giving seven features. The remaining modes are silent. There are zero shared fundamentals, by infrared and Raman mutual exclusion. Counting dimensions gives IR-active, Raman-active and silent vibrational coordinates, totaling .
For the Td molecular point group, the corresponding calculation starts fromand gives . Subtracting translations and rotations leavesThe fundamentals give four IR features. The fundamentals give eight Raman features, of which four coincide with the IR features. The fundamental is silent. The dimensions are .
Thus the predicted IR/Raman/coincidence counts are for the square and for the tetrahedron. These are numbers of distinct ideal fundamental frequencies: each or multiplet contributes one feature, and different copies of the same irreducible representation generally have different frequencies. Accidental degeneracy, overtones and combination bands are outside this count.
Use the printed numbering: site 1 is at the top on , sites 2 and 3 are on the right, site 4 is at the bottom, and sites 5 and 6 are on the left; is perpendicular to the ring. Let be consistently phased, orthonormal atomic orbitals at those sites.
In the Hückel method, and , including the 6–1 bond. Set , with the usual . The cyclic nearest-neighbour matrix has eigenvalues . One real, normalized orthonormal eigenbasis isEach row means . The three lowest molecular orbitals hold the six electrons. Any real orthogonal mixing within either degenerate pair is an equally valid choice of molecular orbitals; the coefficient vectors specify exactly which sketches are used here.
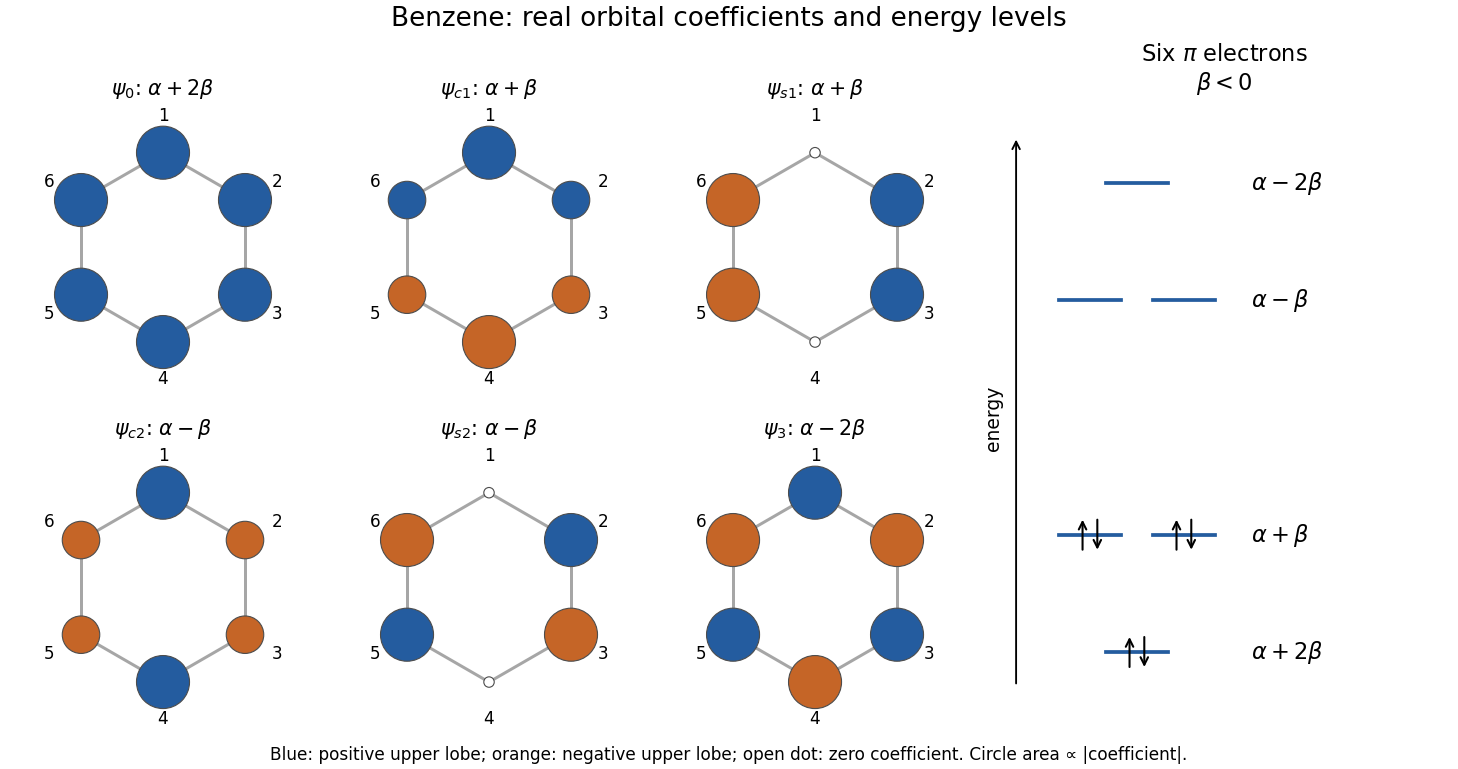
Original benzene pi-orbital energy diagram and six real orbital phase sketches in the printed atom numbering
. Details. 
The coloured circles show the signs and relative magnitudes of the lobes above the molecular plane. The lobes below it have the opposite signs; colours are quantum phases, not electric charges.
Filling the three bonding molecular orbitals with two electrons each givesThree isolated ethene-like bonds, evaluated with the same Hückel method parameters, have . Thus the signed delocalization energy isEquivalently, the positive stabilization is .
Benzene has a planar cyclic conjugated system with six electrons, satisfying the count with . All three bonding molecular orbitals are filled and the antibonding levels are empty, giving the closed-shell stabilization associated with aromaticity. The conclusion uses the continuous cyclic overlap as well as the electron count.
Pyrazine belongs to , the D2h point group. With the printed axes, its three twofold axes are ; it also has an inversion centre and the three coordinate mirror planes. The nitrogen atoms occupy sites 1 and 4 on the axis. Interchanging carbon and nitrogen is not a molecular symmetry operation, so the sixfold benzene symmetry is lost.
The nitrogen pair and the four-carbon set are separately invariant under the D2h point group. Construct symmetry-adapted linear combinations within these two sets; mixing equivalent representations across the sets is left for the Hückel method calculation.
For the nitrogen atomic orbitals, normalized combinations areFor the carbon atomic orbitals, takeThese six functions are orthonormal under the zero-overlap assumption. The suffixes describe coefficient patterns across the carbon sites; the sign acquired by each underlying function must also be included when assigning an irreducible representation. In particular, reflection through the molecular plane changes the sign of every function, while inversion sends to . Ignoring that orbital sign would give incorrect labels.
The complete group representation is thereforeThe diagram shows all six ring symmetry-adapted linear combinations and the resulting energy ordering used in part (iii).
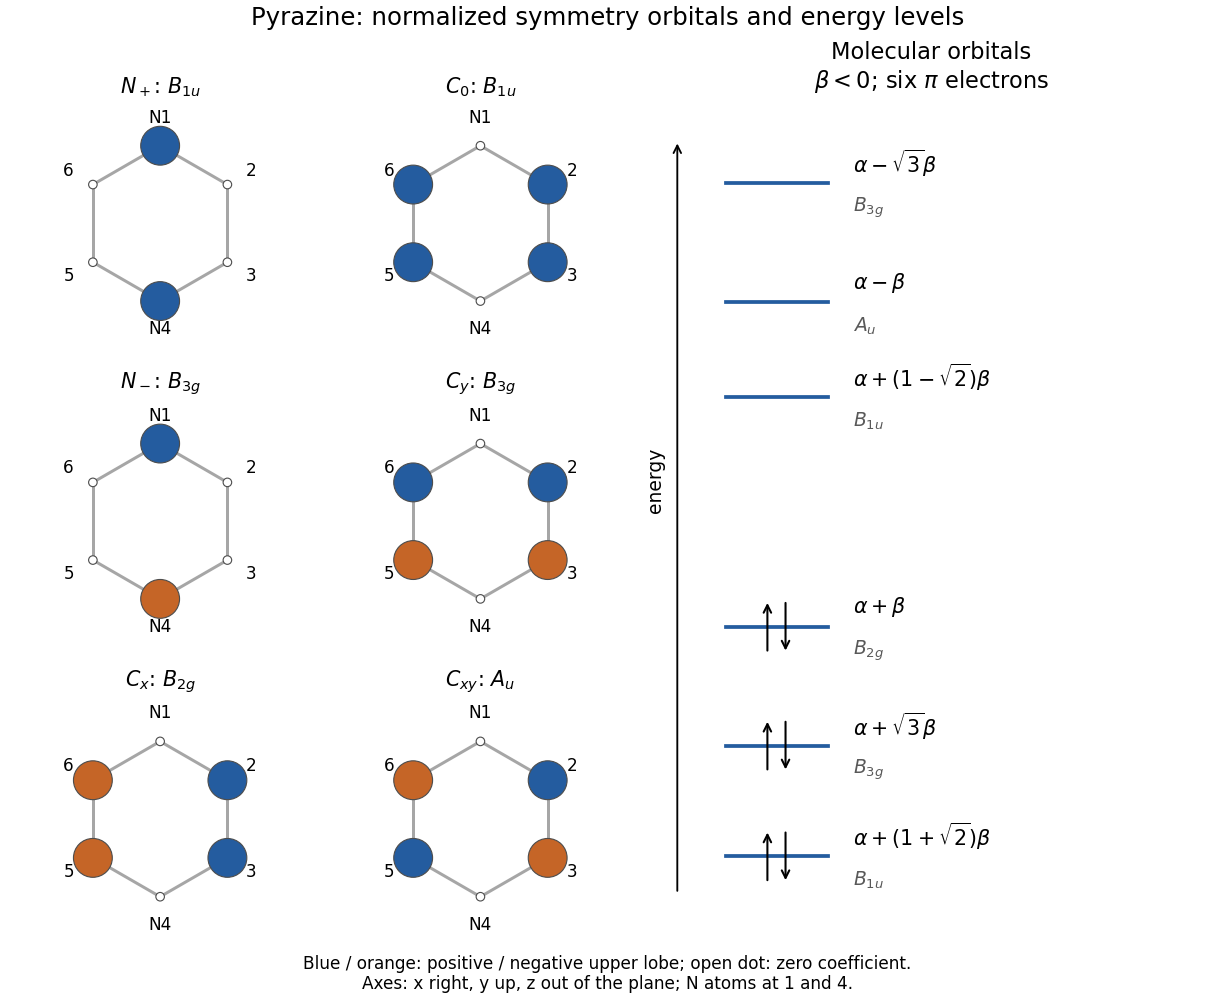
Original normalized pyrazine symmetry-orbital phase patterns and Hückel energy diagram, with nitrogen at sites 1 and 4
. Details. 
Work with . Its nearest-neighbour ring entries are , and its diagonal entries are on the nitrogen sites and on the carbon sites. Because commutes with each molecular symmetry operation, it only mixes symmetry-adapted linear combinations with the same irreducible representation.
In the bases and , the two nontrivial matrix blocks areTheir characteristic polynomials are and . The remaining single-function blocks are and . Thus, ordered from lowest to highest physical energy because , the molecular orbitals areAll six levels are nondegenerate in this parametrization. The right-hand ladder of the diagram in part (ii) shows these levels and occupations. The occupied level is the highest occupied molecular orbital, and the unoccupied level immediately above it is the lowest unoccupied molecular orbital.
The six electrons give the Hückel method energyThe appropriate localized reference contains two C=N bonds and one C=C bond. Three C=C bonds would incorrectly replace the nitrogen one-centre parameters. For one isolated C=N pair, the dimensionless matrix iswhose eigenvalues are . Since , the occupied bonding molecular orbital has energy . Two doubly occupied C=N bonds and one doubly occupied C=C bond consequently giveTherefore the signed delocalization energy isThe positive stabilization is , smaller than the benzene value in the stated model but still stabilizing.
Pyrazine is aromatic: its planar cyclic conjugated system contains six electrons and has a filled set of three bonding molecular orbitals. Each ring nitrogen contributes one electron; its in-plane lone pair is outside this system. The heteroatoms change orbital energies and coefficients without interrupting cyclic conjugation or the count. Aromaticity does not require the stabilization to equal benzene's.
Articles by others on the same topic
There are currently no matching articles.