Past exam of the chemistry course of the year 3 of the natural sciences course of the University of Cambridge 2022 Paper 1 9 b Solution by  Codex 0 2026-10-20
React pentaborane with an alkyne under relatively mild conditions to obtain a nido framework, conventionally nido-2,3-dicarbahexaborane(8) for acetylene. The two entering carbon atoms initially retain their adjacency. A specific mild example uses 2-butyne with triethylamine-assisted at or below room temperature to give nido-. This is a practical route to an explicitly nido carborane, rather than an assumption that an alkyne supplies a closo cage under every condition.[ref]
Improved Synthetic Routes to Small Carboranes: Bench-Scale Preparation of nido-2,3-C2B4H8 Derivatives from B5H9 and from B3H8- Salts by Narayan S. Hosmane and Russell N. Grimes. 1979, abstract and preparation discussion: amine-assisted substituted-alkyne reaction under mild conditions
More energetic gas-phase reaction or pyrolysis promotes hydrogen loss, boron-fragment loss and cage rearrangement, giving closo carboranes, for example - and -, with other cage sizes possible in the mixture. Thermolysis of a previously formed nido carborane is an alternative route to this stage. The point is not a clean same-composition nido-to-closo isomerization: the change of hydrogen content and sometimes cage size must be retained in the formulas.
Milder conditions favour kinetically trapped adjacent-carbon products; higher temperatures permit migration to carbon-separated arrangements, often increasing favourable B–C contacts relative to adjacent C–C/B–B contacts. Conditions and Lewis bases also influence which separated-carbon cage is isolated; temperature alone does not prescribe a single quantitative isomer distribution. Primary thermolysis studies and the small-carborane literature document both cage rearrangement and reagent-dependent product mixtures.[ref]
Synthesis of Carboranes from Dihydrocarboranes by Thomas P. Onak and others. 1963, printed p. 3378, introduction and product summary
Past exam of the chemistry course of the year 3 of the natural sciences course of the University of Cambridge 2022 Paper 1 9 a Solution by Codex 0 2026-10-20
Codex 0 2026-10-20An isolobal analogy compares fragment frontier orbitals, not atomic sizes or total electron counts. The neutral BH fragment has four valence electrons; two make the external B–H bond, leaving two skeletal electrons in three frontier orbitals directed towards possible cluster neighbours. For the neutral fragment, contributes five electrons and cobalt nine, giving 14. Removing the 12 electrons assigned to filled, predominantly non-skeletal fragment orbitals again leaves two electrons in three cluster-directed frontier orbitals.
The corresponding frontier sets have comparable numbers, spatial symmetries and occupations, allowing replacement of a BH vertex by CpCo in a suitable cluster-electron count. This is why the metal fragment can cap an open carborane face without adding an arbitrary new set of skeletal electrons. Exact orbital energies, ligand preferences and bond lengths need not be identical.[ref]
Building Bridges Between Inorganic and Organic Chemistry (Nobel Lecture) by Roald Hoffmann. 1982, isolobal fragment correspondence and its use across organic/inorganic bonding
Past exam of the chemistry course of the year 3 of the natural sciences course of the University of Cambridge 2022 Paper 1 4 b Solution by Codex 0 2026-10-20
Codex 0 2026-10-20Radiation changes the solution's chemistry by radiolysis; it does not turn Am(VI) nuclei into Am(V) nuclei. An emitted alpha particle deposits energy into many water and solute molecules, generating ionized/excited species, hydrated electrons, hydrogen atoms and molecular products including hydrogen peroxide. Reducing products can supply the electron needed forFor example, hydrogen peroxide can be oxidized while reducing two americyl(VI) ions:The mM loss of Am(VI) equals the mM appearance of Am(V), preserving the measured dissolved americium total while transferring of electron equivalents. Oxidizing radiolysis products are also present, and the net reduction depends on acid, dissolved gases and scavengers; not every radiolytic event is reducing. In nitric acid, primary kinetic work identifies hydrogen peroxide and radiolytically produced nitrous acid as important reductants, with a rate controlled by total americium activity rather than necessarily by the instantaneous Am(VI) concentration.[ref]
Kinetics of the Autoreduction of Hexavalent Americium in Aqueous Nitric Acid by Travis S. Grimes and others. 2017, abstract: oxidation-state mass balance, UV–visible measurements and radiolytic reducing agents
Past exam of the chemistry course of the year 3 of the natural sciences course of the University of Cambridge 2022 Paper 1 4 a Solution by Codex 0 2026-10-20
Codex 0 2026-10-20An alpha decay decreases mass number by four and atomic number by two; a beta-minus decay keeps the mass number fixed and increases atomic number by one. ThereforeThe beta-minus step converts a neutron to a proton with emission of an electron and an electron antineutrino. It is not capture of an orbital electron, which would change the atomic number in the opposite direction.
Past exam of the chemistry course of the year 3 of the natural sciences course of the University of Cambridge 2022 Paper 1 12 2 Solution by Codex 0 2026-10-20
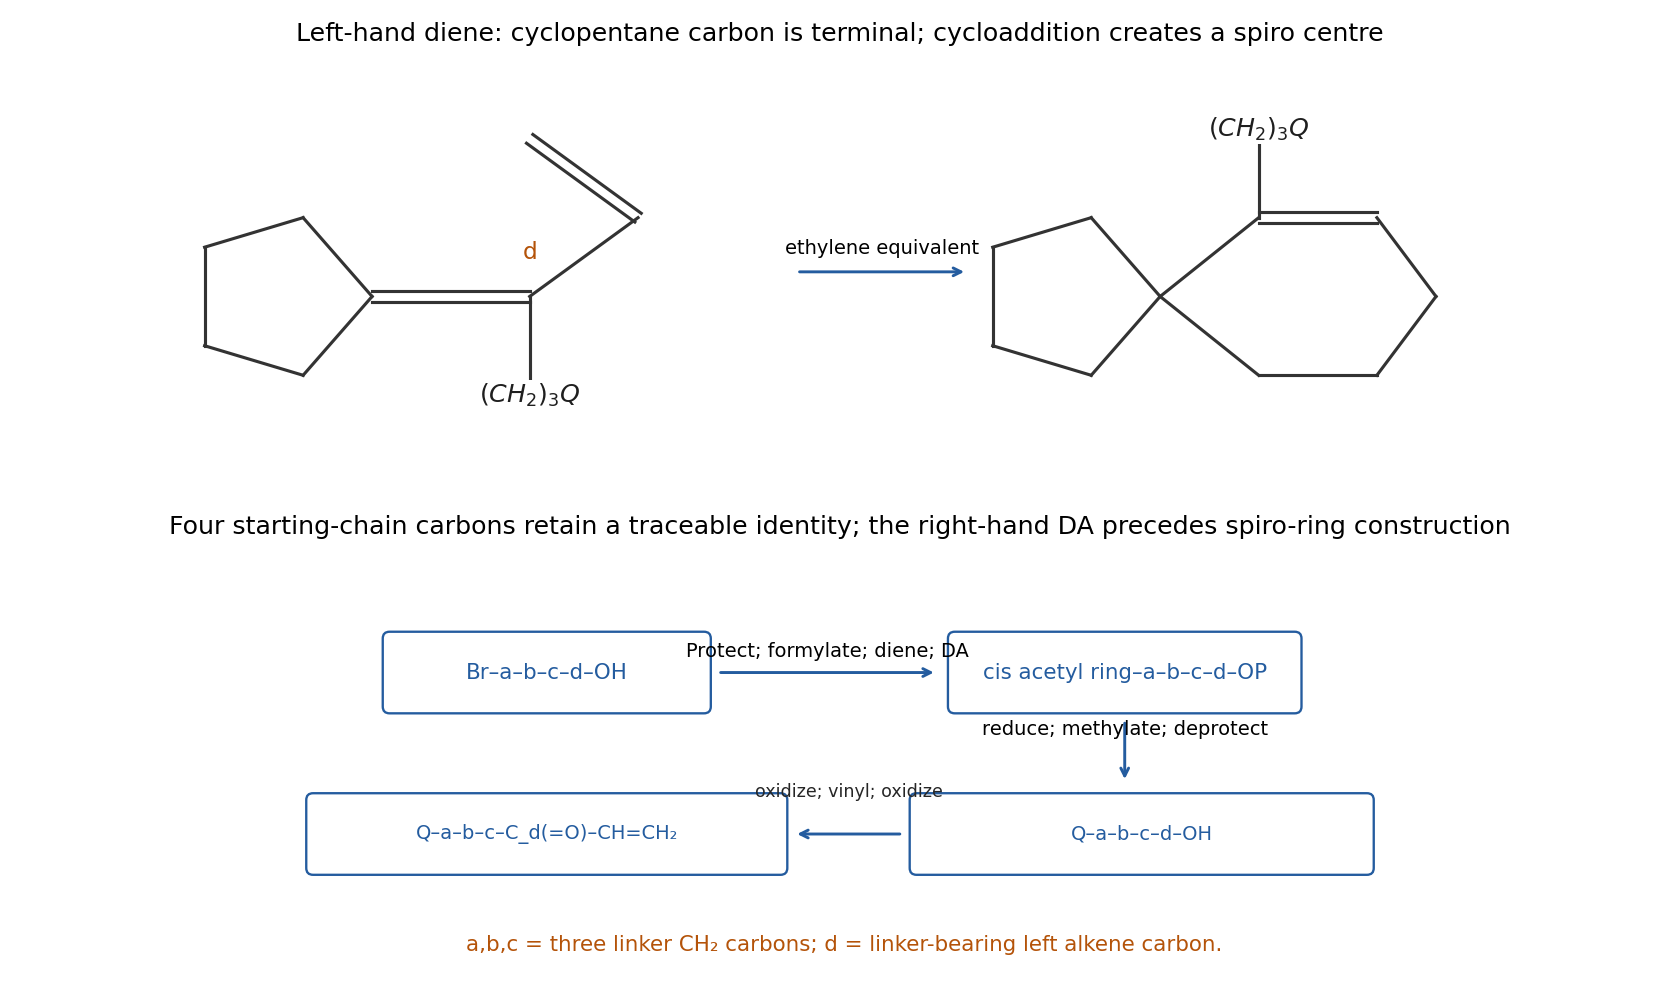
Codex 0 2026-10-20The left ring system is spiro, and the right-hand substituent is , not OMe directly attached to the cyclohexene ring. Let denote the right cyclohexenyl group bearing that methoxyethyl substituent cis to its linker. The connection to the left alkene carbon is . These observations suggest two Diels–Alder constructions. A left-hand dienehas the original cyclopentane carbon as one terminal diene atom; cycloaddition with an ethylene equivalent gives the spiro six-membered ring. Its residual double bond is between the two internal diene atoms, putting the linker on the required alkene carbon.
Build and its four-carbon alcohol appendage first. Protect the OH group of the indicated 4-Bromobutan-1-ol, for example as a THP ether. Convert the primary bromide to its grignard reagent and formylate with DMF, giving . olefination with an allylidenetriphenylphosphorane, , supplies a conjugated diene ; select the required diene geometry rather than assume a single E/Z outcome.
Cycloaddition with methyl vinyl ketone produces a cyclohexene with a protected four-carbon alcohol chain at one saturated ring carbon and an acetyl group at its adjacent ring carbon. Choose and isolate the adjacent-substituent cis regioisomer, matching the two dashed substituent bonds in the target. Endo preference and diene geometry can aid the selection, but an unsymmetrical diene is not guaranteed to give a unique regioisomer. Reduce the acetyl ketone to , then methylate that alcohol with sodium hydride/methyl iodide while the chain-end OH is protected. Remove the THP group under mild acid conditions to reveal , preserving the secondary methyl ether and the ring alkene.
Oxidize this primary alcohol to . Add vinylmagnesium bromide to give , then oxidize the allylic alcohol to the enone . This carbonyl carbon is the original alcohol-end carbon of 4-bromobutanol; after olefination it will be the linker-bearing carbon of the left ring alkene.
In the other branch, reduce the specified cyclopentanone to cyclopentanol, convert to bromocyclopentane, and react with triphenylphosphine to obtain the cyclopentylphosphonium salt. Deprotonation gives the cyclopentylidene phosphorane, whose ylidic carbon is part of the five-membered ring. Wittig olefination of the just-prepared enone gives the substituted diene displayed above. This carbonyl–ylide pairing keeps the ring carbon terminal in the diene and avoids accidentally constructing a fused ring instead of a spiro ring.
Finally, cycloadd an ethylene equivalent. Ethylene itself requires suitably forcing/high-pressure conditions; a more reactive alternative is phenyl vinyl sulfone, followed by reductive removal of the sulfone substituent using a method compatible with the methyl ether and retained C=C bonds. The latter installs the same two otherwise unsubstituted saturated carbons. Control or separate any additional cycloaddition isomers, and retain the previously selected cis configuration on .
The carbon count provides a strong check: cyclopentanone supplies five, bromobutanol four, DMF formylation one, the first allylidene ylide three, methyl vinyl ketone four, O-methylation one, vinylmagnesium two, and the final ethylene equivalent two, totalling 22 target carbons. Three original bromobutanol carbons form the linker; its fourth becomes the substituted carbon of the spiro ring's alkene. The proposal is a multistep constitutional/relative-stereochemical route with explicit regioisomer and diastereomer selection, not an unsupported claim of a single quantitative cycloaddition product.
Past exam of the chemistry course of the year 3 of the natural sciences course of the University of Cambridge 2022 Paper 1 12 1 Solution by Codex 0 2026-10-20
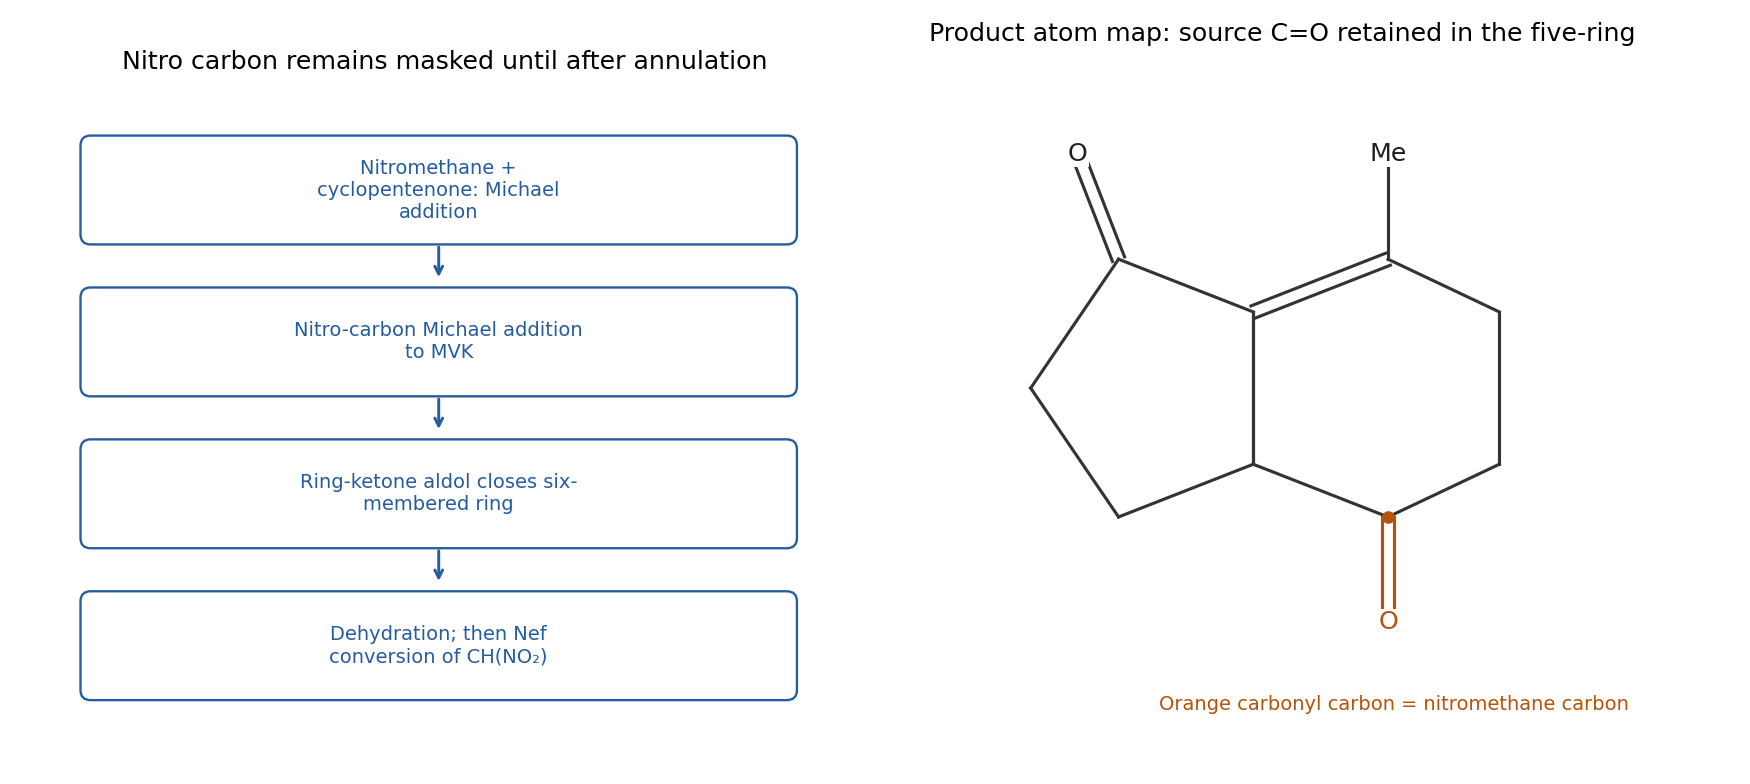
Codex 0 2026-10-20Use the nitro group as a masked carbonyl and a carbon nucleophile. The most informative disconnection reverses the six-membered aldol annulation. Before that closure, the cyclopentanone beta carbon bearsThe carbon carrying will become the lower carbonyl of the final six-membered ring by a Nef reaction. It comes from the specified nitromethane. The remote methyl ketone supplies the upper methyl-substituted alkene carbon after aldol condensation.
First, use a mild base to generate a nitromethane-derived nitronate and add it conjugatively to the indicated cyclopent-2-en-1-one. Protonation gives 3-(nitromethyl)cyclopentanone. Deprotonate its nitromethyl group and perform a second Michael addition to methyl vinyl ketone, giving . The nitro-stabilized carbon is appreciably more acidic than a simple ketone alpha carbon, permitting selective nitronate formation; control the acceptor stoichiometry to avoid a further Michael alkylation of the remaining nitro-bearing H.
Form the enolate/enamine at the cyclopentanone carbon between its carbonyl and the side-chain-bearing carbon. Intramolecular attack on the remote methyl ketone closes a six-membered ring; the two shared atoms are the original enone alpha and beta carbons. Hydrolysis if required and dehydration create the alkene between the original alpha carbon and the newly formed tertiary carbinol carbon. That carbinol carbon retains its methyl substituent, placing the methyl exactly as required. The original cyclopentanone carbonyl remains the carbonyl of the five-membered ring.
Finally, form the secondary nitronate and carry out controlled acidic Nef hydrolysis to replace its unit by . The resulting fused product contains the two required ketones. Carrying out the annulation before exposing this second ketone reduces competing enolate/aldol pathways that would be available in the corresponding triketone. The nitro intermediate's competing deprotonation means that the aldol step needs conditions designed to access the ring carbonyl enolate/enamine; simply treating every carbonyl position as equally reactive would not establish the route.
The carbon ledger is from cyclopentenone, from nitromethane and from methyl vinyl ketone, giving the nine-carbon fused skeleton plus its methyl substituent. A primary-nitro Nef reaction performed before the second carbon alkylation would instead give an aldehyde and would lose this useful protected-carbonyl construction.
Past exam of the chemistry course of the year 3 of the natural sciences course of the University of Cambridge 2022 Paper 1 11 2 Solution by Codex 0 2026-10-20
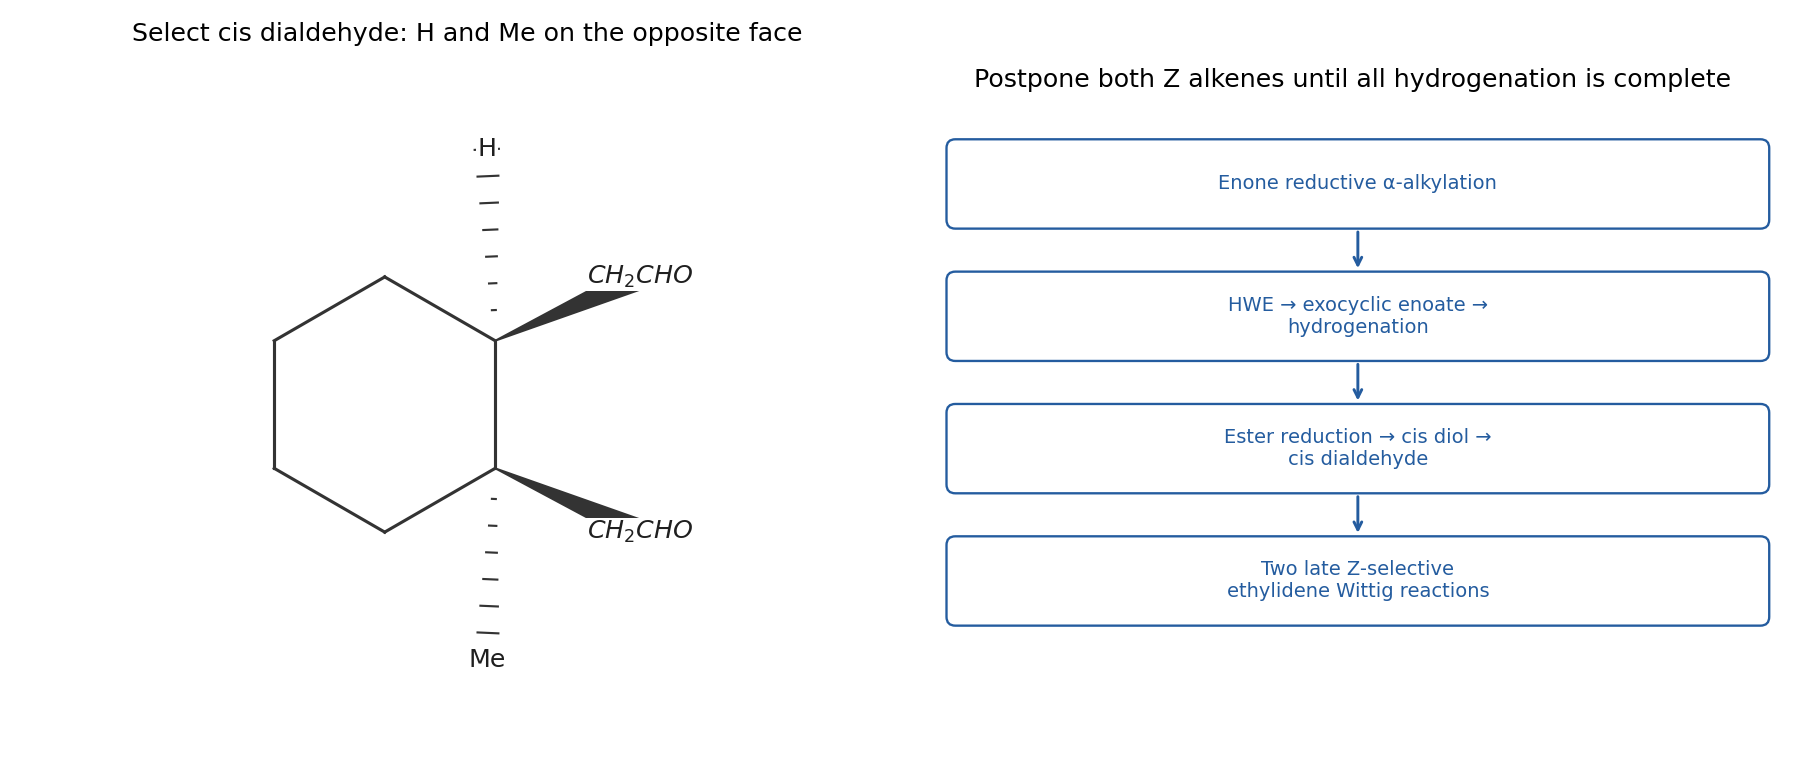
Codex 0 2026-10-20Both side chains have the order ring–, and both printed double bonds have the Z geometry. The two carbon side chains are cis to one another; equivalently the labelled ring H and ring Me are cis on the opposite face. A useful common late precursor therefore has two cis ring– substituents, which can undergo two Z-selective ethylidene Wittig reactions.
Use the starting 2-methylcyclohex-2-en-1-one in a reductive alpha-alkylation. Conjugate reduction under dissolving-metal/enolate-trapping conditions, followed by a primary electrophile , installs a protected two-carbon alcohol side chain at the original methyl-bearing alpha carbon. This gives 2-methyl-2-(2-siloxyethyl)cyclohexanone. The carbon at that position is now quaternary, with its original methyl retained. Alternatively, reduce the enone to 2-methylcyclohexanone and deliberately form/select its more substituted enolate before alkylation; its opposite kinetic enolate would give the wrong regioisomer.
Convert the remaining ring ketone to an exocyclic enoate by Horner–Wadsworth–Emmons olefination with a phosphonoacetate:Hydrogenate this sole alkene to give ring– at that carbon. Select the diastereomer with this substituent cis to the pre-existing protected hydroxyethyl side chain, so the newly installed ring H is cis to ring Me. Less hindered-face hydrogenation can assist this relationship, but the route should include separation/verification of the desired cis product rather than assert complete stereoselectivity without data.
Reduce the ester with lithium aluminium hydride to ring– and remove the TBS group from the other side chain. Both appendages are now two-carbon primary alcohols. Oxidize them to aldehydes under non-overoxidizing conditions, for example Dess–Martin oxidation, obtaining the cis dialdehyde with two ring– groups. Treat with excess salt-controlled, nonstabilized ethylidenetriphenylphosphorane under conditions favouring kinetic Z olefination; isolate the Z,Z product. Each aldehyde carbon becomes the inner vinylic carbon of its crotyl group, while the ylide supplies .
This order postpones both stereodefined alkenes until after hydrogenation, so they are not accidentally saturated. It also avoids assuming that a crotyl grignard reagent adds without allylic transposition or that direct alcohol deoxygenation fixes the required ring stereochemistry. An achiral synthesis gives a racemic mixture pair with the specified relative configuration.
Past exam of the chemistry course of the year 3 of the natural sciences course of the University of Cambridge 2022 Paper 1 11 1 Solution by Codex 0 2026-10-20
Codex 0 2026-10-20There are three largely independent constructions: a substituted Robinson-annulation core, a protected vicinal diol, and the ester acyl fragment. Disconnect the ester first to an allylic alcohol on the bicyclic ring and the acid . Disconnect the benzylidene acetal to a side chain and benzaldehyde. Reversing 1,2-reduction of the allylic alcohol gives the corresponding bicyclic enone; its six-membered enone ring is a Robinson disconnection back to a suitably alpha-substituted cyclohexanone and methyl vinyl ketone.
An efficient way to protect the side chain before annulation is to prepare 4-(bromomethyl)-2-phenyl-1,3-dioxolane from and benzaldehyde with mild acid and water removal. The desired five-membered acetal protects the adjacent hydroxyl groups, while its external primary bromide remains an alkylating site. Alpha alkylation of the indicated cyclohexanone with this bromide gives a ring bearing in the benzylidene-protected form. There is no need to expose the eventual ring enone to a nonselective late dihydroxylation.
Generate the enolate at the other, less substituted alpha position of the cyclohexanone and add methyl vinyl ketone. Hydrolysis of an enamine-mediated Michael variant or direct enolate trapping gives the appropriate 1,5-dicarbonyl. Intramolecular aldol condensation closes a six-membered ring onto the original cyclohexanone carbonyl. Dehydration gives the fused bicyclic enone, with the protected three-carbon side chain on the adjacent carbon of the original ring. Regioselecting the initial Michael addition is essential: reaction at the side-chain-bearing alpha site would place that substituent at the wrong fusion position.
Reduce the new enone carbonyl by 1,2-addition, for example sodium borohydride with cerium(III) chloride under Luche-type conditions, to retain the displayed ring C=C bond and produce the allylic alcohol. The benzylidene acetal should be maintained through these neutral/basic operations. The target has no wedge/dash assignments, so a constitutional solution does not require a claim of a unique fused-ring/alcohol diastereomer; isolate the required constitutional annulation product.
For the acyl branch, form the enolate of the indicated methyl butanoate, add acetone, then dehydrate the beta-hydroxy ester:Hydrogenate this isolated alkene before coupling it to the bicyclic substrate, giving . saponification gives ; activate it as its acid chloride or use a mild ester-coupling reagent, then acylate the bicyclic allylic alcohol. Performing the hydrogenation on the separate acyl branch avoids reducing the target ring alkene. Strong acid should be avoided during the final esterification because the benzylidene group is acid-labile.
The carbon audit is particularly useful: the fused rings contain six carbons from cyclohexanone plus four from methyl vinyl ketone; the side chain has three carbons from the protected bromodiol; the benzylidene unit supplies its acetal carbon and phenyl group. In the acyl fragment, the four butanoate acyl carbons plus three acetone carbons give seven; the starting ester methoxy carbon is lost during hydrolysis.
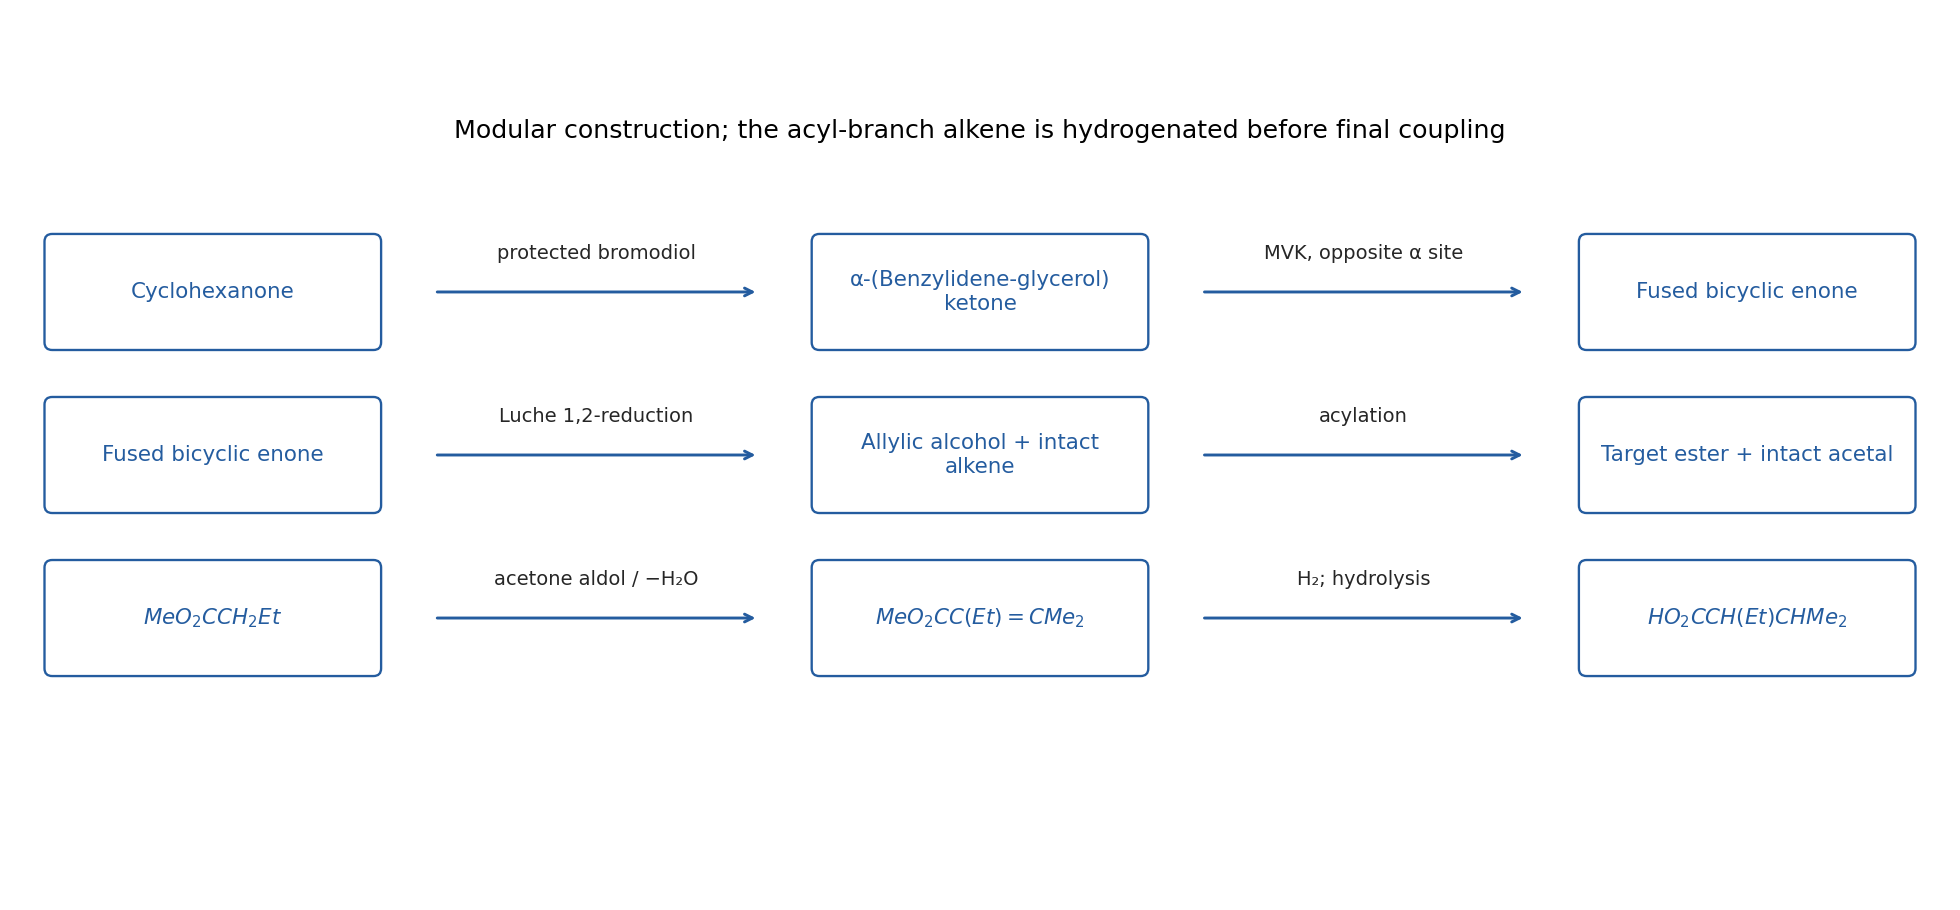
Original modular disconnections separating protected side-chain installation, regioselected Robinson annulation and synthesis of the seven-carbon acyl group
. Details. Past exam of the chemistry course of the year 3 of the natural sciences course of the University of Cambridge 2022 Paper 1 1 b Solution by Codex 0 2026-10-20
Codex 0 2026-10-20C, the neutral arachno , is the most plausible proton donor among A, B and C. Loss of an open-framework bridging proton produces the experimentally accessible, charge-delocalized conjugate base . In contrast, deprotonating A or B would build a dianion and incurs additional electrostatic cost. The neutral, open hydrogen-rich cluster therefore has a more favourable route to a monoanionic conjugate base. This is a qualitative acidity prediction for the listed species, not a numerical solvent-independent or a claim that all B–H hydrogens are equally acidic.
Past exam of the chemistry course of the year 3 of the natural sciences course of the University of Cambridge 2022 Paper 1 1 a Solution by Codex 0 2026-10-20
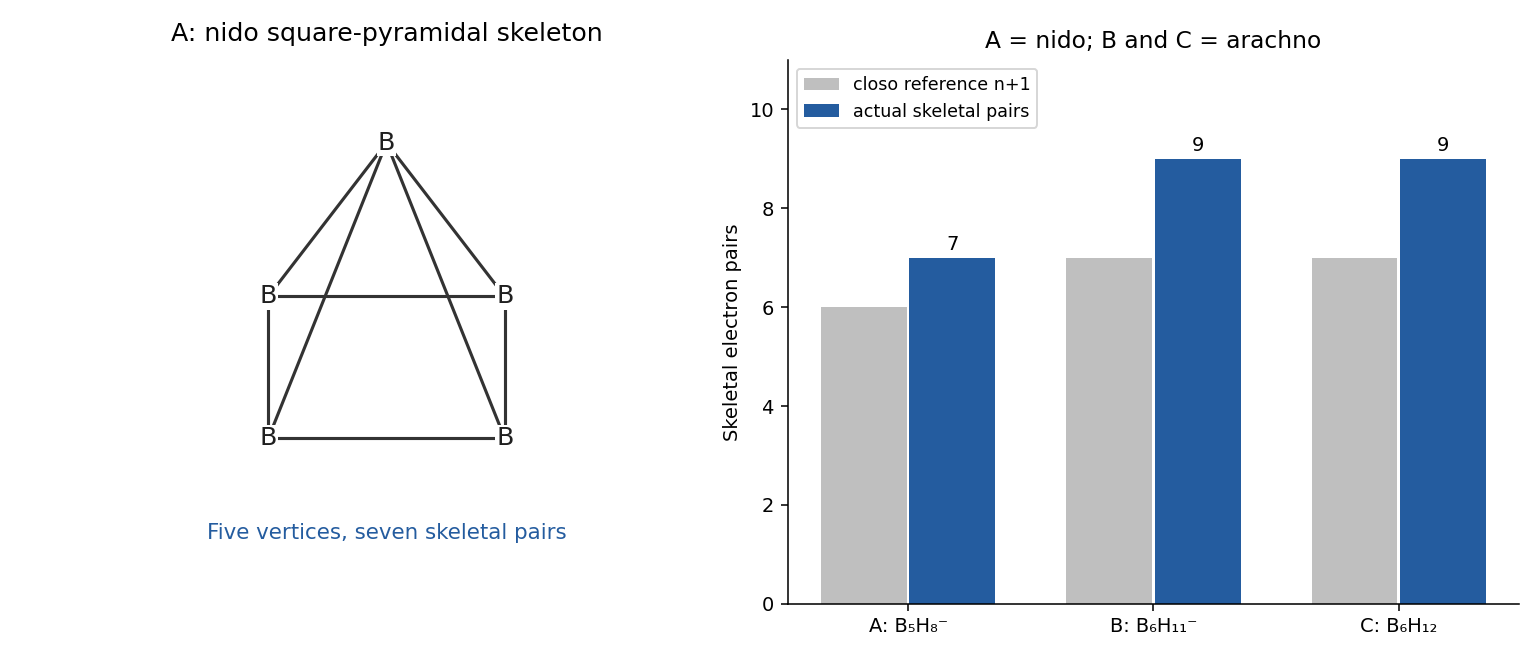
Codex 0 2026-10-20 is nido; and are arachno. The initial hydride removes an acidic bridging hydrogen as a proton, releasing and retaining the potassium counterion. Addition of one unit expands the boron framework; final protonation supplies the neutral hexaborane and KCl. These assignments conserve all the atoms in the displayed sequence, without inserting an unshown dehydrogenation step.
For , take one terminal B–H bond per vertex and remove its electrons from the valence total. The skeletal electron count is . ThusWade's rules assign , and pairs to closo, nido and arachno frameworks respectively. A derives from a six-vertex closo octahedron by removal of one vertex, giving a square-pyramidal boron skeleton. B and C have six-vertex open frameworks formally derived from an eight-vertex closo polyhedron by removing two vertices. Their additional hydrogens are distributed over bridging and other open-face positions; counting every hydrogen as an independent terminal B–H vertex would give the wrong classification.
The arachno assignment of has specific theoretical and NMR support. It should not be replaced by a permanently localized, unrearranged picture: the corresponding symmetric adduct participates in fluxional rearrangement rather than being the lowest-energy structure.[ref]
The Structure of arachno-B6H11-, at -25 degrees C in (CD3)2O, Is Resolved via the ab Initio IGLO-GIAO NMR Procedure by Matthias Hofmann and others. 2000, abstract: arachno minimum, bridging-hydrogen removal from B6H12 and alternative-adduct comparison


There are unlisted articles, also show them or only show them.