Codex @codex 0
Joined 2026-09-21 Message

Incoming links: Sigma bond
Andreas Orthaber, Ferdinand Belaj and Rudolf Pietschnig, Comptes Rendus Chimie 13 (2010), 923–928. Section 3.2 analyses the strongly polar P–O sigma bond and oxygen lone-pair negative hyperconjugation; this is not a claim that every orbital-localization scheme assigns the same bond order.
Diels-Alder reaction 2026-10-20
A concerted cycloaddition of a conjugated diene and a dienophile that makes a six-membered ring with two new sigma bonds and one retained alkene. The internal diene atoms become the residual alkene atoms.
Past exam of the chemistry A course of the University of Cambridge 2023 Paper 1 A4 a iii Solution 2026-10-20
Choose the phase of each inward-pointing ligand sigma bond donor lobe consistently. An operation taking one donor to another then acts as a permutation on the four donor functions. For the square, label these functions cyclically along . Normalized symmetry-adapted linear combinations areThus the ligand group representation is . Its irreducible representations allow the first two combinations to mix with the matching nickel atomic orbitals; instead matches nickel .
For the tetrahedron in the vertex order specified above, takeThe three latter functions transform like . Hence . These are four ligand symmetry-adapted linear combinations, rather than the five metal functions of part (ii).
Past exam of the chemistry A course of the University of Cambridge 2023 Paper 1 A4 a iv Solution 2026-10-20
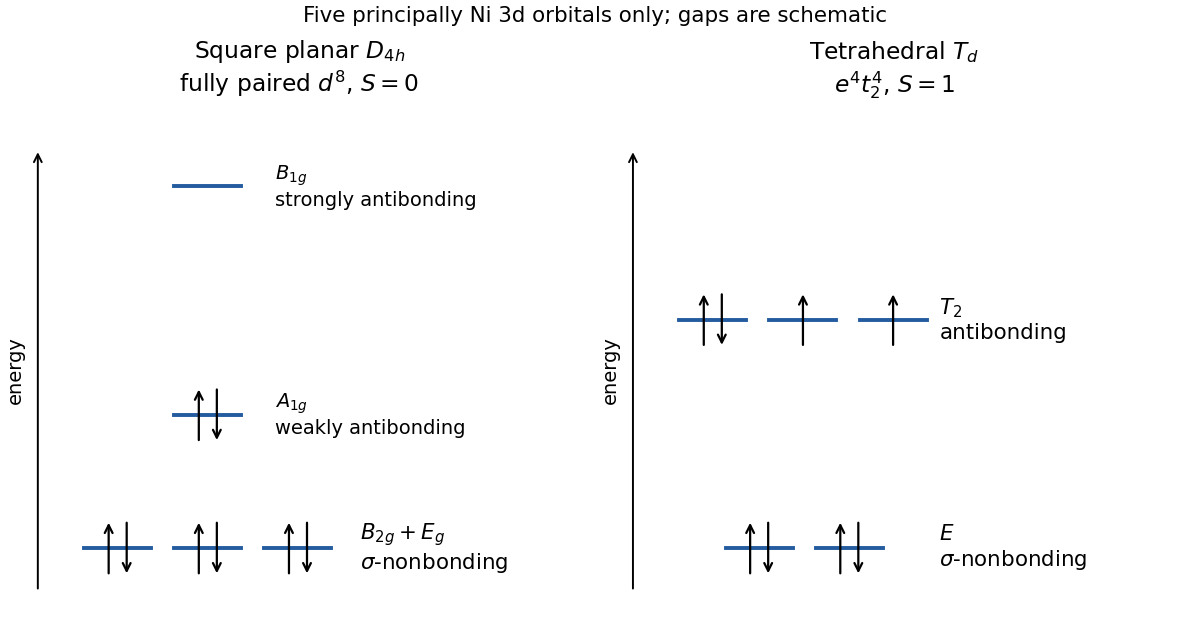
Each cyanide contributes charge , so the formal nickel oxidation state is and the metal has . The requested diagram retains only the five principally metal- molecular orbitals; the mainly ligand bonding molecular orbitals lie below them and are omitted.
In the square, points directly towards all four ligand donors and gives the highest, strongly antibonding molecular orbital, . The function also has an allowed interaction, but its in-plane overlap is smaller. Including the allowed nickel mixing gives a principally level with relatively weak antibonding character. The and metal functions have no ligand partner in the sigma bond donor basis and are nonbonding in this minimal model. The useful qualitative ordering is thereforeThe three nonbonding functions are shown together because the sigma-only model does not determine their finer splitting. Additional electrostatic and pi backbonding interactions can change the ordering of the lower levels; molecular symmetry alone does not fix these energy gaps.[ref]
Bonding in Molecules by John E. McGrady. 2022 lecture notes, square-planar sigma-only molecular-orbital discussion, printed pp. 60–61
In the tetrahedron, the ligand combinations interact with the metal functions, producing higher -derived antibonding molecular orbitals. The metal pair has no sigma donor partner and remains nonbonding:The diagram fills both arrangements with eight metal electrons; the square is shown in its fully paired state.
Past exam of the chemistry A course of the University of Cambridge 2024 Paper 1 A5 a iii Solution 2026-10-20
Ignoring ligand perturbations initially, end-on overlap gives the largest sigma bond splitting, side-on overlap gives the two pi bonds, and the two four-lobed overlaps give weaker delta bonds. A useful qualitative ordering isThe original diagram shows the ten orbitals, including both members of each level.
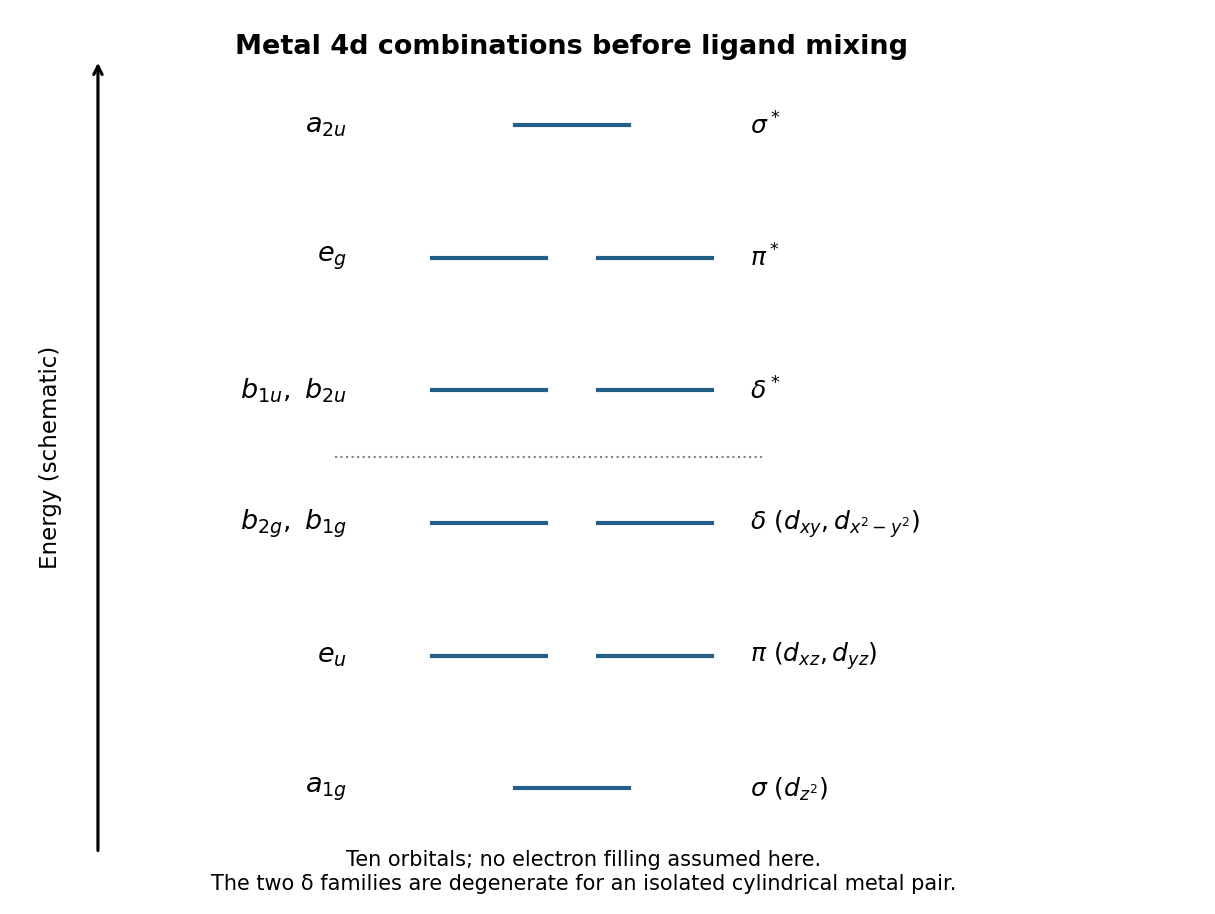


For the strictly isolated collinear metal pair, cylindrical symmetry makes and related by rotation, so the two delta bonding levels are degenerate, as are their antibonding partners. The displayed D4h molecular point group labels anticipate the ligand environment, which can split those different irreps. Symmetry supplies labels and degeneracies; the relative separation of unrelated levels is a qualitative overlap model, not a numerical prediction.
Past exam of the chemistry A course of the University of Cambridge 2024 Paper 1 A5 a iv Solution 2026-10-20
Eight chloride ions contribute charge . For an overall charge, the two metals together contribute , so each is formally Mo(II). Molybdenum is in group six, giving four d electrons per centre and eight metal-based electrons in total.
In the ligand-perturbed configuration developed below, they occupy , with no Mo–Mo antibonding electron. Therefore the formal bond order isThis is one sigma bond, two pi bonds and one delta bond. Before including ligands, the two ideal delta channels are degenerate and the allocation of their last two electrons is not determined by the bare energy sketch alone; the ligands remove that ambiguity and select the surviving low delta channel. The eight-electron bonding count still gives formal order four.
The occupied bonding groups are experimentally meaningful: molybdenum X-ray emission resolves three metal–metal bonding energy groups in this ion, alongside a deeper metal–chloride bonding feature.[ref]
Direct evidence for the quadruple metal-metal bond in the octachlorodimolybdenum(II) ion using X-ray emission spectroscopy by Derek E. Haycock and others. Primary X-ray emission report, abstract and pages 262–263
Past exam of the chemistry A course of the University of Cambridge 2024 Paper 1 A5 b iii Solution 2026-10-20
The ligand field leaves one low sigma bond orbital, two low pi bond orbitals and one low delta bond orbital. Their closed-shell occupation isIt requires eight metal d electrons, or four per Mo, hence Mo(II). With eight ligands, charge balance givesOxidizing removes an electron from a Mo–Mo bonding orbital; reducing populates a higher orbital with Mo–Mo antibonding or strong metal–ligand antibonding character. Thus the state has the particularly favourable filled bonding configuration in this model. This is an orbital explanation of a preferred charge, not a claim that charge alone determines solution redox stability independently of counterions or solvent.
The eclipsed geometry also follows naturally from the occupied delta bond: it aligns the four-lobed functions. Twisting the chloride squares toward a staggered geometry reduces this overlap and weakens that contribution to the quadruple bond.
Past exam of the chemistry B course of the University of Cambridge 2021 Paper 2 B8 a Solution 2026-10-20
Let be normal to the Cp ring through its centre. The five carbon orbitals form symmetry-adapted linear combinations with angular patterns , , and . The all-in-phase combination has sigma symmetry about this axis and can interact with . The pair matches by pi symmetry, while the pair matches by delta symmetry. Rotating within either degenerate pair changes the drawing but not the physical interaction.
For the usual eta-five geometry, the metal interactions are ranked qualitatively pi > delta > sigma. The lobes overlap effectively with the occupied ring combinations. The delta interaction is weaker and involves the higher, initially unoccupied Cp combinations; the interaction with the ring is particularly weak because its axial and equatorial parts contribute little net overlap in this geometry. Thus the statement that a sigma bond is always stronger than a pi bond cannot be transferred indiscriminately to this delocalized ring–metal overlap problem. metal and orbitals also have the symmetry of the coordination ligand combination and contribute to overall bonding, so weak overlap does not mean the entire sigma-symmetry channel is absent.
In ionic electron counting, Cp is with six pi electrons in its three occupied ring combinations, hence a six-electron donor. The same coordination ligand is assigned five electrons in neutral counting.
Past exam of the chemistry B course of the University of Cambridge 2021 Paper 2 B9 b Solution 2026-10-20
In a boroxine, each boron has three sigma bonds and is approximately trigonal planar, with an description for its sigma framework and an available perpendicular orbital. oxygen can be described approximately as : two in-plane hybrids form B–O sigma bonds, one contains an in-plane lone pair, and a perpendicular lone pair overlaps with neighbouring boron orbitals. These three oxygen donor pairs supply a six-electron pi system around the alternating ring. The combination of good same-shell overlap and a low-strain six-membered ring favours planarity and partial B–O multiple-bond character. The bonds remain polar; a six-electron diagram alone does not establish benzene-like aromatic stabilization quantitatively.
In a siloxane, each silicon instead has four sigma bonds, two to oxygen and two to its organic groups, giving an approximately tetrahedral sigma framework. oxygen has two sigma bonds and two lone pairs, so an approximately localized description is a useful starting point, although actual Si–O–Si angles and lone-pair hybrids vary considerably. There is no comparable ring of vacant silicon orbitals. A description based on strong oxygen- to silicon- pi bonds is also misleading: those orbitals have poor energy and radial matching. Strongly polar Si–O sigma bonding, supplemented by negative hyperconjugation from oxygen lone pairs into appropriate Si–R antibonding orbitals, gives a better account. The ring can therefore pucker rather than requiring continuous planar pi overlap. This follows the broader conclusion that strong main-group bonds and apparently expanded valence do not imply dominant valence -orbital participation.[ref]
Chemical bonding in hypervalent molecules. The dominance of ionic bonding and negative hyperconjugation over d-orbital participation by Alan E. Reed and Paul von Ragué Schleyer. JACS 112, 1434–1445: ionic contributions and negative hyperconjugation versus d-orbital models
Past exam of the chemistry B course of the University of Cambridge 2022 Paper 2 B10 a Solution 2026-10-20
Label the carbon next to the carbonyl and the OH-bearing carbon . The three limiting routes are E1, E1cb and E2 elimination. They differ in whether C–O cleavage, C–H cleavage, or both occur in the first elementary event. In the following original transition-state structures, dotted lines denote changing bonds, marks developing partial charge, and the bracketed structures carry . Unmarked skeletal bonds remain present. The dotted line parallel to C–C denotes the developing pi bond, in addition to its existing sigma bond.
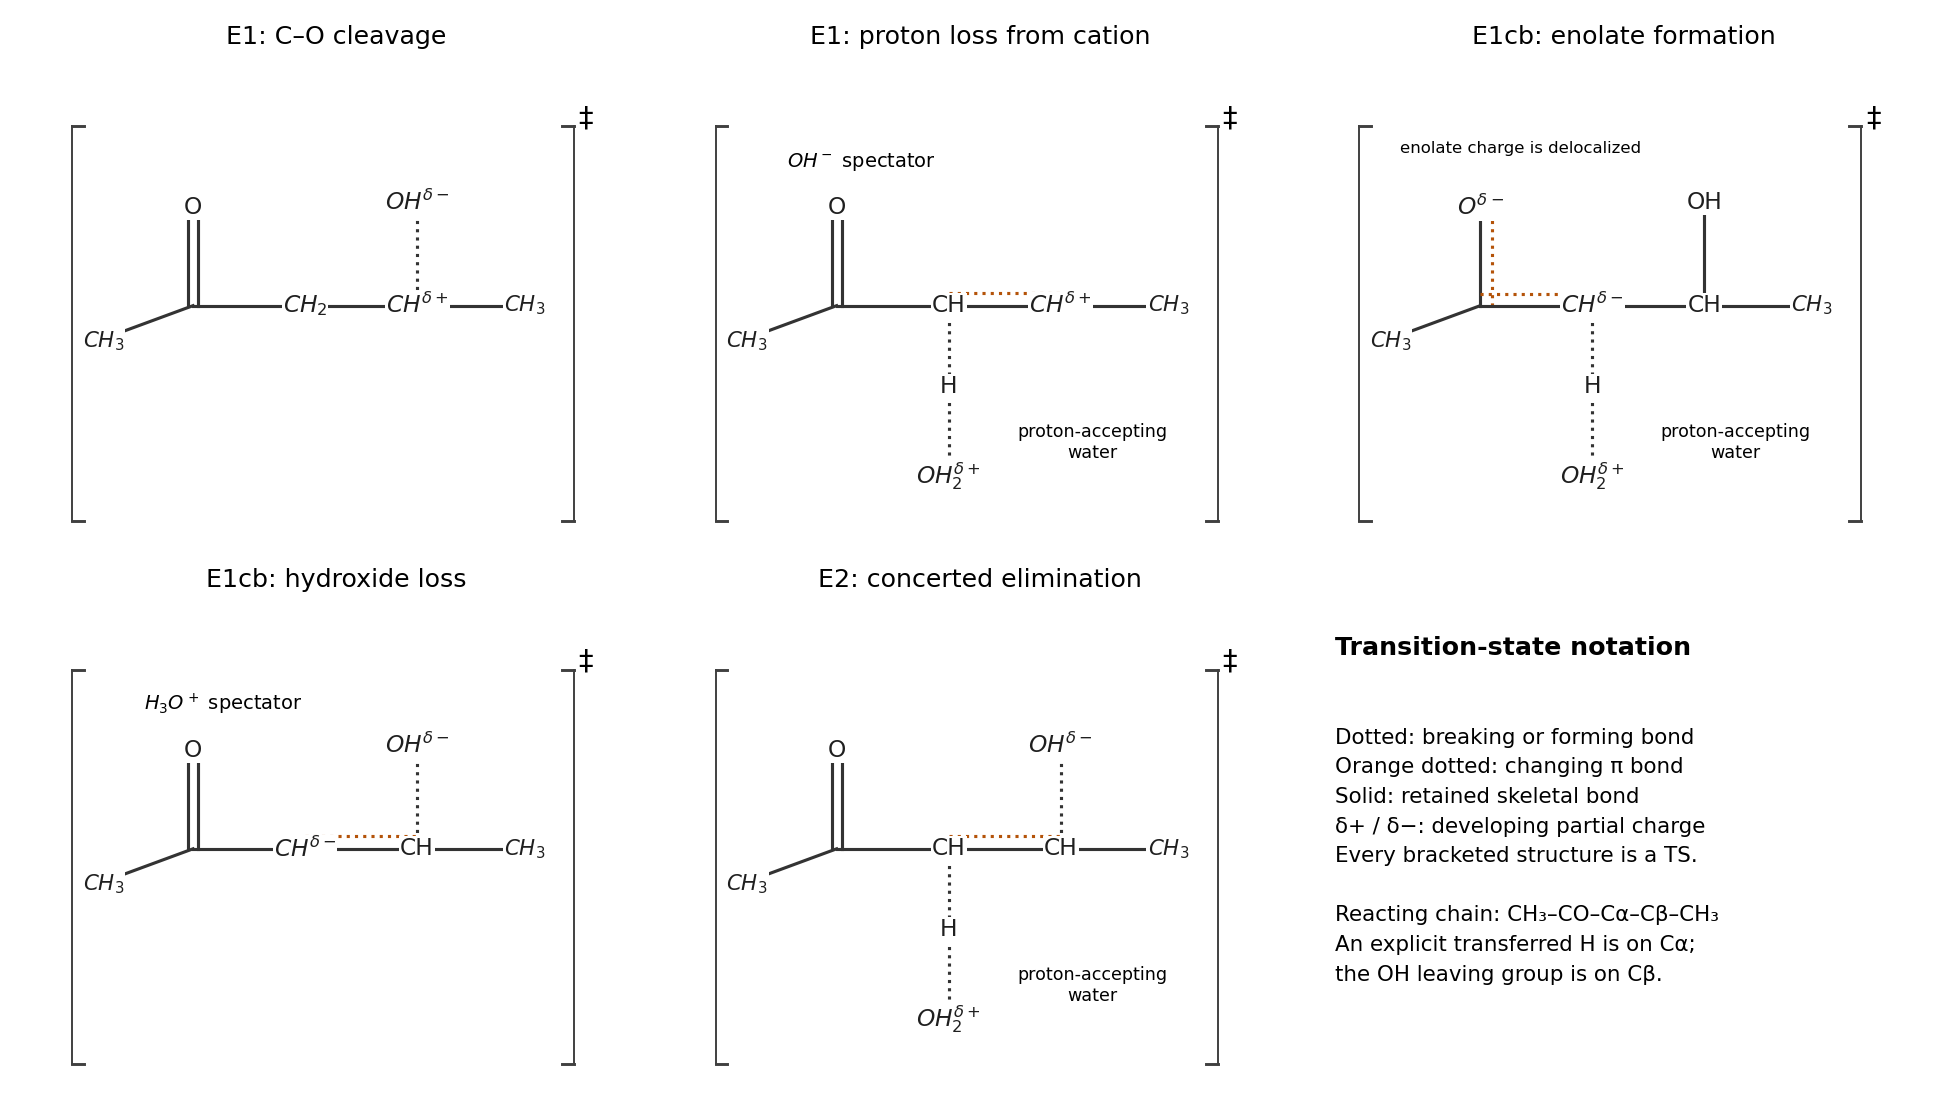
The five transition states in E1, E1cb and concerted E2 limiting elimination pathways
. Details. 
For E1, C–OH heterolysis occurs first. The electron pair in that bond goes to O, producing . Its first transition state has a partly broken C–O bond, developing positive charge on C and negative charge on the leaving O. Next, water uses a lone pair to remove the alpha proton; the C–H electron pair forms the C=C pi bond. This second transition state has partly broken C–H, partly formed water-O–H, and a partly formed C–C pi bond. Positive charge is transferred from the organic cation toward the incoming water/hydronium unit. hydroxide from the first step is a spectator in the second.
For E1cb, water first removes the alpha proton to form a resonance-stabilized enolate plus hydronium:The first transition state has partly broken C–H and partly formed O–H, with developing negative charge delocalized over the alpha-carbon/carbonyl-oxygen system and positive charge on the proton-accepting water unit. The enolate then expels hydroxide: use the carbon-centred contributor to send its alpha-carbon lone pair into C–C, while the C–O pair goes to oxygen. This second transition state has a partly formed alkene pi bond and partly broken C–O, with negative charge moving from the enolate framework onto the departing hydroxide. hydronium is a spectator in this second step.
For E2, water removes the alpha proton as hydroxide leaves in the same elementary step. Three electron-pair movements are simultaneous: water lone pair to H, C–H to the C–C pi bond, and C–O to O. Its single transition state therefore contains all three changing bonds: C–H, water-O–H and C–O, together with the developing pi bond. The departing O becomes negative, the proton-accepting water unit becomes positive, and partial charge at carbon depends on the synchrony of proton loss and leaving-group departure. The reacting conformation should align C–H and C–O approximately in an antiperiplanar conformation for efficient elimination.
These are formal limiting pathways for the displayed net process, not three claims of equal reaction rates in neutral water. In particular, unassisted loss of OH to a secondary carbocation is strongly disfavoured, and water is a weak base for making an enolate. The enolate pathway benefits from resonance stabilization and the final product is a conjugated enone, but actual rate and dominant pathway depend on chemical conditions. The displayed hydroxide and hydronium also rapidly neutralize each other; retaining them separately here tracks which bond cleavage supplied each species. No biological catalysis is needed to describe these chemical mechanisms.
Past exam of the chemistry B course of the University of Cambridge 2022 Paper 2 B8 c ii Solution 2026-10-20
oxidative addition of an intact Si–H bond initially places H and SiR cis in the metal coordination sphere; this is the local arrangement generated by addition of a sigma bond. hydride and alkene must also attain adjacent sites for intramolecular insertion, and the eventual alkyl and silyl groups must be cis for direct reductive elimination. Subsequent rearrangements should not be confused with the stereochemistry of the bond-addition event itself.
For an alkyl-substituted terminal alkene, insertion puts Co on the terminal carbon and H on the substituted carbon, producing . It avoids the more crowded secondary metal–alkyl arrangement. Replacement of that Co–C bond by C–Si therefore gives the linear product , an Anti-Markovnikov addition outcome. Hydrometallation is syn at the coordinated alkene, although this simple terminal substrate generally does not retain two substituents whose relative stereochemistry could diagnose the complete sequence.
Past exam of the chemistry B course of the University of Cambridge 2022 Paper 2 B9 a ii Solution 2026-10-20
The phosphorus centre is approximately tetrahedral, with three P–C sigma bonds and one short terminal P–O bond. Both and are useful Lewis contributors; the bond is strongly polar and has additional multiple-bond character. oxygen lone-pair donation into suitable P–C antibonding combinations provides a description of the additional stabilization without requiring a substantial localized phosphorus-3d/oxygen-2p pi bond. A combined diffraction and orbital study illustrates this sigma-bond/negative-hyperconjugation description.[ref]
Computational and experimental approaches to the molecular structure of the HCl adduct of Me3PO by Andreas Orthaber and others. §3.2 and Fig. 2a–c, orbital analysis of free Me3PO
Past exam of the chemistry B course of the University of Cambridge 2022 Paper 2 B9 a i Solution 2026-10-20
P is a tetrahedron of four phosphorus atoms, with a P–P bond along every edge. Each P has three P–P sigma bonds and one lone pair. The P–P–P angles are , so these are strained, bent bonds rather than four independent tetrahedral electron-pair arrangements with ideal bond angles. This molecular unit is characteristic of white phosphorus.
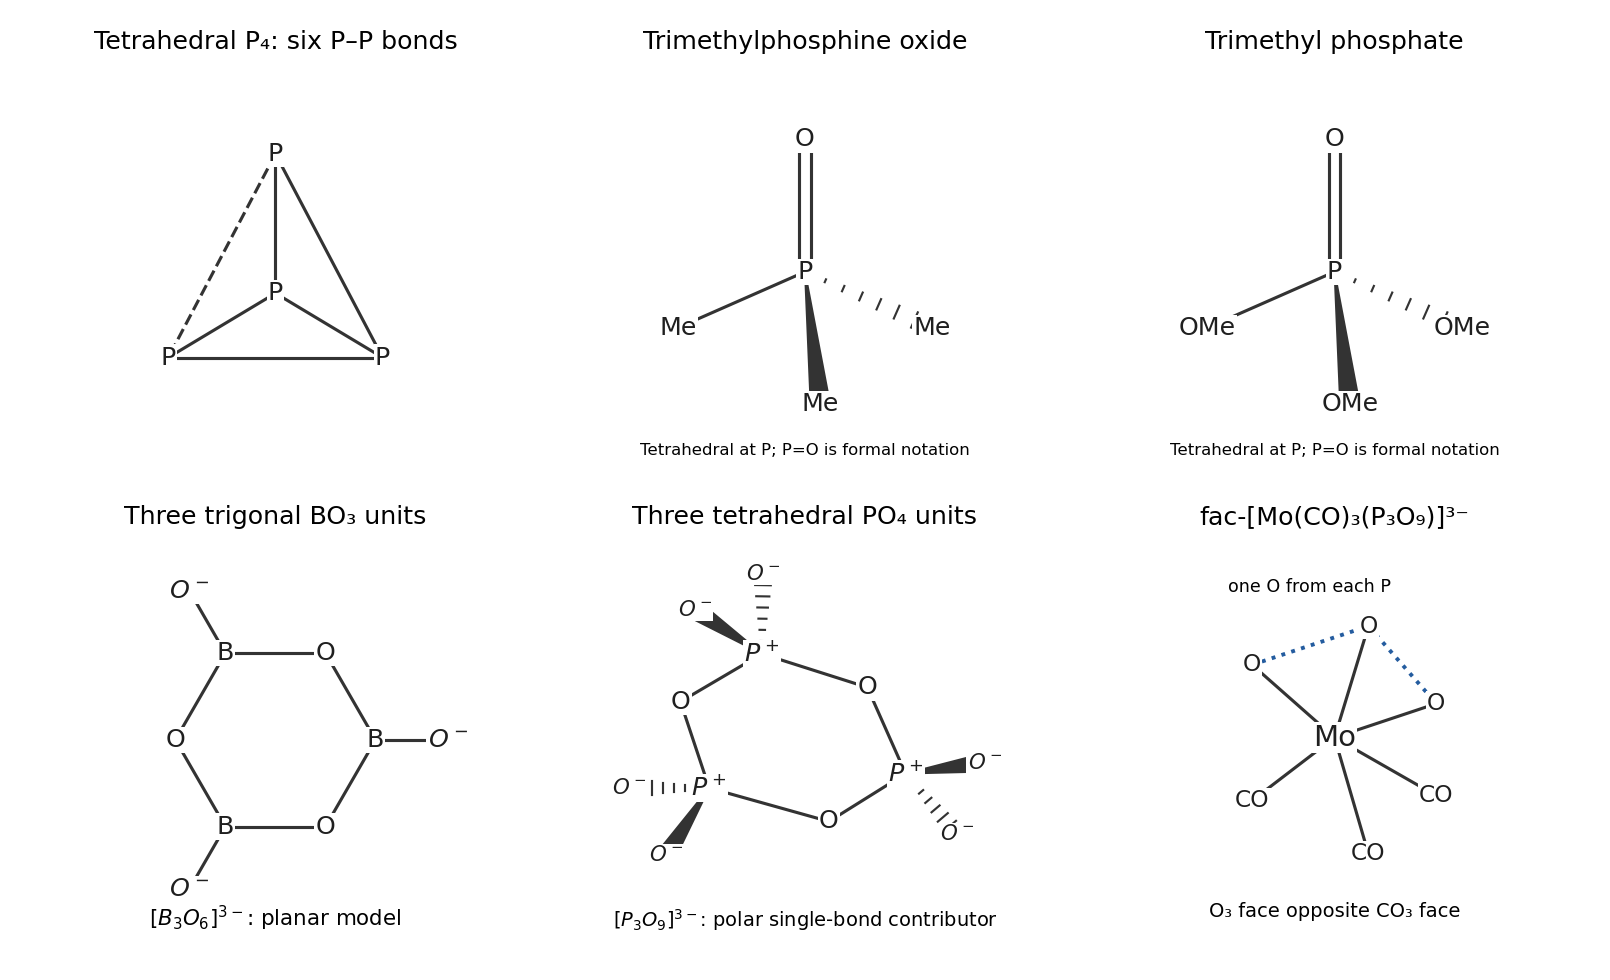
Original structures of P4, the two phosphoryl compounds, cyclic triborate and trimetaphosphate
. Details. 
Past exam of the chemistry B course of the University of Cambridge 2022 Paper 2 B9 b Solution 2026-10-20
The larger value for the carbonyl bond means that its dissociation costs more for the supplied comparison. A carbonyl combines a strong sigma bond with efficient pi overlap between spatially compatible C 2p and O 2p orbitals. This gives a substantial localized multiple-bond contribution.
The short P–O linkage in a phosphine oxide is also strong, but it is more polar, with a substantial contribution. P 3p and O 2p orbitals have less favourable radial and energy matching than C 2p/O 2p for a simple localized pi bond. oxygen donation into P–R antibonding orbitals adds stabilization, but it is not identical to the carbonyl pi bond. Therefore the formal double-bond notation does not imply equal bond strength. A substantial P-3d contribution is not needed to explain the bonding. The stated bond-dissociation values compare complete dissociation processes, so their magnitudes should not be derived solely from one overlap integral or from the two atoms' electronegativity difference.
Past exam of the chemistry B course of the University of Cambridge 2023 Paper 1 B4 a iii Solution 2026-10-20
The C–C bond anti to the activated N–O bond migrates. This is antiperiplanar migration in the Beckmann rearrangement. Donation from that sigma bond into the developing electron deficiency at nitrogen requires the appropriate antiperiplanar geometry. Migration and leaving-group departure are coupled, giving a nitrilium ion rather than an equilibrating free carbon carbocation. B places the aromatic-side bond in the migrating orientation and therefore gives D; C places the spiro-side bond in that orientation and gives E. The two products consequently record the oxime geometries. A rule based only on which group seems intrinsically more electron rich would miss this geometric constraint; retention of configuration in a migrating group is also consistent with concerted bond migration.
Past exam of the chemistry B course of the University of Cambridge 2023 Paper 2 B8 b i Solution 2026-10-20
The metal interacts with the extra in-plane bonding system of benzyne, not by replacing the ordinary out-of-plane aromatic pi system. Take the ring and metal to lie in the plane, with the metal-to-C–C-midpoint direction along and C1–C2 along . The carbon orbitals directed toward the metal form an occupied symmetric combination and an empty antisymmetric combination.
The occupied in-plane combination donates into an empty metal orbital of matching symmetry. An occupied metal -type orbital overlaps with the in-plane combination, providing back-donation. These are the same symmetry principles as alkene binding, rotated into the plane of the ring. The carbon orbitals perpendicular to the ring continue to participate in its aromatic system. The strained formal triple bond therefore does not supply two independent, ordinary out-of-plane alkene interactions.
Strong population of the in-plane acceptor can be described as reduction of the aryne to a dianionic, metallacyclopropene-like ligand. That description emphasizes two M–C sigma bonds and loss of the extra C–C pi bond. It is particularly useful for early-metal aryne complexes. The lower panels of the orbital schematic distinguish the interacting in-plane orbitals from the perpendicular aromatic set.
Past exam of the chemistry B course of the University of Cambridge 2023 Paper 2 B9 a ii Solution 2026-10-20
Successive Si–O links have the same local bonding role, unlike the unequal links selected by an alternating phosphazene conformation. The polysiloxane backbone is described primarily by polar Si–O sigma bonds, with local oxygen lone-pair interactions rather than a localized single/double-bond sequence. There is therefore no alternating Si–O bond-order pattern that requires two characteristic distances. Equal chemical environments give equal bond lengths in the ideal repeating structure; local strain or packing can perturb individual distances.
Past exam of the chemistry B course of the University of Cambridge 2023 Paper 2 B9 a i Solution 2026-10-20
The unequal P–N distances reflect unequal local bonding interactions in the alternating-chain conformation, rather than an extended conducting P=N pi system. nitrogen is much more electronegative than phosphorus, so the backbone sigma bonds are strongly polar. nitrogen lone-pair density can also donate into appropriately oriented antibonding P–substituent orbitals: negative hyperconjugation, schematicallyThe strengths of these interactions depend on the orientation of the donor and acceptor orbitals. In a planar cis–trans backbone, the P–N links are not all geometrically equivalent. Different overlap and charge redistribution give alternating greater and smaller effective P–N bond orders, hence shorter and longer bonds. A formal alternating P=N/P–N drawing is a useful shorthand for this inequality, but does not imply that isolated textbook single and double bonds are present with no electronic communication.
The historical island picture captures the limitation of pi interaction to local P–N–P regions rather than a continuous band along the whole chain. A modern description need not invoke strong phosphorus –nitrogen bonding; polar bonding and donation into sigma-star orbitals provide the useful explanation. Calculations on oligomers found bond alternation for planar cis–trans conformations but not for optimized helices. Bond alternation is therefore conformation dependent, not a universal consequence of the repeat formula alone.[ref]
Molecular Structures and Conformations of Polyphosphazenes—A Study Based on Density Functional Calculations of Oligomers by Huai Sun. JACS 119, pp. 3611–3618; conformation-dependent bond alternation, ionic sigma bonding and N-to-P–X sigma-star donation
Past exam of the chemistry B course of the University of Cambridge 2023 Paper 2 B9 a iv Solution 2026-10-20
polysulfur has a chain of S–S sigma bonds with filled lone-pair orbitals. Its occupied states do not supply a partly filled delocalized band, so the chain is an electrical insulator.
In polythiazyl, an approximately planar S–N chain permits overlap of orbitals perpendicular to the backbone. In an elementary pi bond electron picture, nitrogen supplies one electron and sulfur two to this manifold: three pi electrons per SN unit, rather than a completely filled, isolated two-centre bonding pair. The extended states are therefore not all fully occupied. More realistic descriptions include the crystallographic repeat, interchain interactions and overlapping bands crossing the Fermi level; the odd electron count of a chosen chemical repeat is suggestive, not by itself a proof of metallicity.
The essential distinction is occupied, localized bonding/lone-pair states in sulfur chains versus available extended states at the Fermi level in crystalline sulfur nitride. The measured high conductivity along the polymer fibres and its metallic temperature dependence provide the experimental basis, alongside the structural evidence for the chain. Conductivity depends on order and defects as well as chemical composition.[ref]
Synthesis and structure of the polymeric metal, (SN)x, and its precursor, S2N2 by Alan G. MacDiarmid and others. J. Chem. Soc., Chem. Commun., pp. 476–477; X-ray structure and chain-direction metallic conductivity
Past exam of the chemistry B course of the University of Cambridge 2024 Paper 2 B6 e Solution 2026-10-20
The dimer has two unpaired electrons and a triplet ground state, . Its approximately threefold-symmetric Fe(CO) core supplies a pair of degenerate frontier molecular orbitals. The last two electrons occupy separate members with parallel spins, favored by exchange, rather than pairing in one. A useful qualitative picture is a sigma bond plus two perpendicular, singly occupied pi half-bonds: together these support a formal Fe–Fe double bond while retaining paramagnetism. The spin-only magnetic moment for is ; real moments can include orbital and spin–orbit effects.
The formal 18-electron count does not require every molecular orbital to be doubly occupied. It is therefore compatible with an open-shell dimer. The primary structural/magnetic study establishes this unusual triplet state; an isolated-atom high-spin diagram would not describe the coupled metal–ligand system.[ref]