Past exam of the chemistry B course of the University of Cambridge 2024 Paper 1 B4 d Solution by  Codex 0 2026-10-20
The free propargylic OH first reacts with an aluminium hydride, releasing hydrogen and forming an O–Al bond. Coordination of the nearby alkyne to this aluminium center organizes a five-membered O–C–C–C–Al unit. Hydride addition from the opposite side of the activated alkyne gives a cyclic alkenyl aluminate: aluminium bonds to the distal alkyne carbon, while hydride is delivered to the carbon nearer .
The key step is directed trans hydroalumination. The new C–H and C–Al bonds are anti. Aqueous workup protonolyzes C–Al with retention of its alkene geometry and hydrolyzes the O–Al attachment. Replacing that C–Al bond by C–H therefore produces the E allylic alcohol, with and the rest of the chain on opposite alkene sides. A generic syn hydroalumination drawing followed by simple protonation would predict the wrong geometry.
The free OH provides the directing tether; the silyl-protected oxygen is not equivalently available. Primary electrophile-trapping studies of Red-Al-treated propargylic alcohols support the alkenyl-alanate description.[ref]
trans-Hydroalumination/Alkylation: One-Pot Synthesis of Trisubstituted Allylic Alcohols by Neil F. Langille and Timothy F. Jamison. 2006, pp. 3761–3764, trans-hydroalumination and trapping of alkenyl alanates
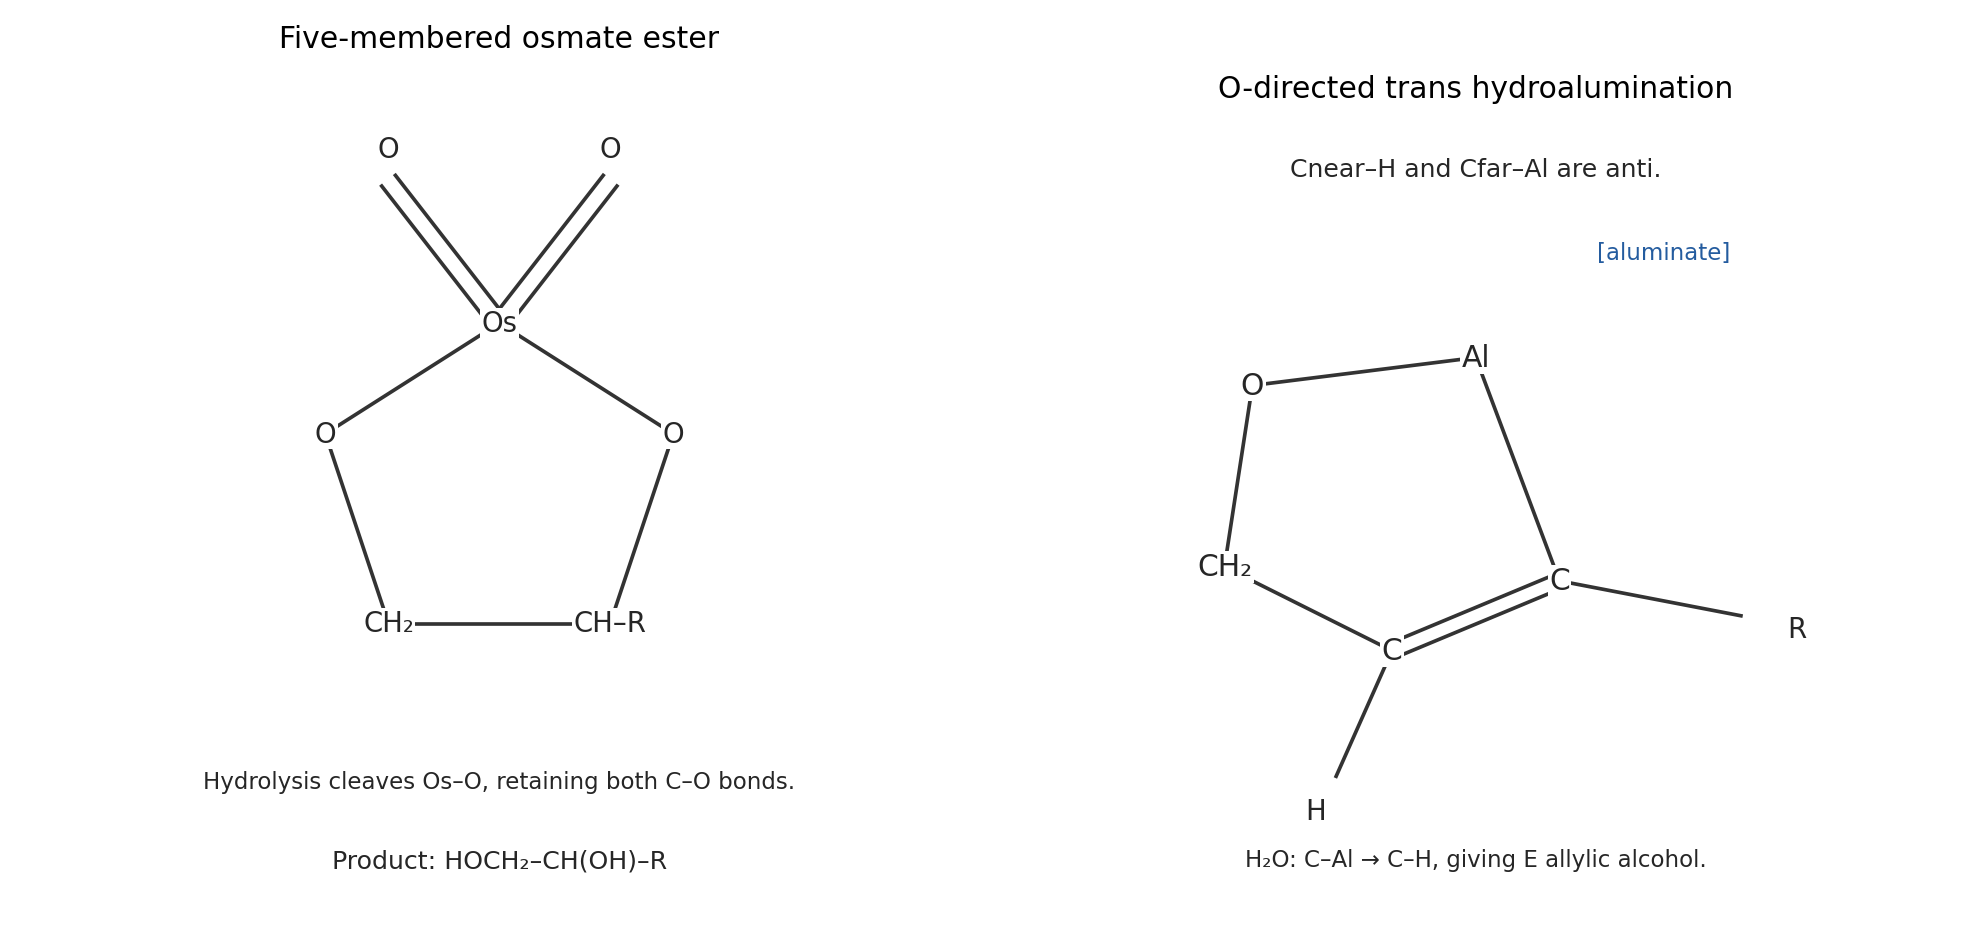
Cyclic intermediates in osmium dihydroxylation and directed trans hydroalumination
. Details. Past exam of the chemistry B course of the University of Cambridge 2024 Paper 1 B4 c Solution by Codex 0 2026-10-20
Codex 0 2026-10-20The reacting alkene pi bond and two Os=O units undergo a concerted cycloaddition to give a five-membered cyclic osmate ester. Both new carbon–oxygen bonds form on the same face. Hydrolysis and proton transfer cleave the Os–O attachments, releasing the vicinal diol and a reduced osmium-containing species.
This is syn dihydroxylation of the terminal alkene. The terminal carbon becomes , and the adjacent carbon becomes ; the remote vinyl chloride remains as drawn. The protected region denoted by is a spectator for this local mechanism. No explanation of alkene chemoselectivity is required here. Since no terminal reoxidant is specified, do not invent a catalytic regeneration step for Os(VIII); osmium reduction and product release suffice for the stated transformation.
Past exam of the chemistry B course of the University of Cambridge 2024 Paper 1 B4 b Solution by Codex 0 2026-10-20
Codex 0 2026-10-20Acid protonates one methoxy group of 2,2-Dimethoxypropane. methanol departure gives the oxocarbenium species , which is captured by one diol oxygen. After proton transfer, protonation and loss of the remaining methoxy group generate a second oxocarbenium center. Intramolecular attack by the other diol oxygen closes the five-membered ring; deprotonation regenerates acid. The net transacetalization exchanges two methoxy groups for the two diol oxygens, releasing two methanol molecules.
The two starred carbon centers retain their original configurations, and the two carbon side chains on the acetonide are trans. No C–O bond at either original stereocenter is broken; reaction takes place at the reagent-derived CMe acetal carbon. The apparent change in wedge/dash presentation comes from redrawing the backbone into the five-membered ring, not inversion at a starred center. In the planar pentagon used in the diagram, draw the alkynyl-bearing side chain behind and the side chain in front. The unstarred methyl-bearing center also retains its configuration. The acetal carbon itself is not stereogenic because its two methyl substituents are identical.
Past exam of the chemistry B course of the University of Cambridge 2024 Paper 1 B4 a Solution by Codex 0 2026-10-20
Codex 0 2026-10-20The alkene undergoes concerted oxygen transfer from the terminal peroxide oxygen of mCPBA. In the cyclic electron-flow description, the alkene pi bond begins one C–O bond, the peroxide oxygen supplies the second, O–O breaks, and proton transfer/carboxyl rearrangement gives m-chlorobenzoic acid. Both C–O bonds are installed from the same alkene face, so the alkene's relative substituent geometry is retained in the epoxide. There is no freely rotating carbocation intermediate.
The neighboring free OH hydrogen-bonds to the approaching peracid and favors a cyclic, organized transition-state geometry. oxygen is delivered from the OH-directed face, giving the indicated syn relationship between epoxide oxygen and the allylic hydroxyl. In the supplied projection, the epoxide C–O bonds and the original C–OH bond are on the rear/hashed face. Steric interactions still influence the preferred hydrogen-bonded conformer, but “attack from the least hindered face” alone misses the positive directing role of OH. The distant benzyl ether is not the directing free hydroxyl.
The classical experimental comparison of allylic alcohols and protected derivatives established this directing principle; it supports the mechanistic interpretation without itself measuring the selectivity of this acyclic substrate.[ref]
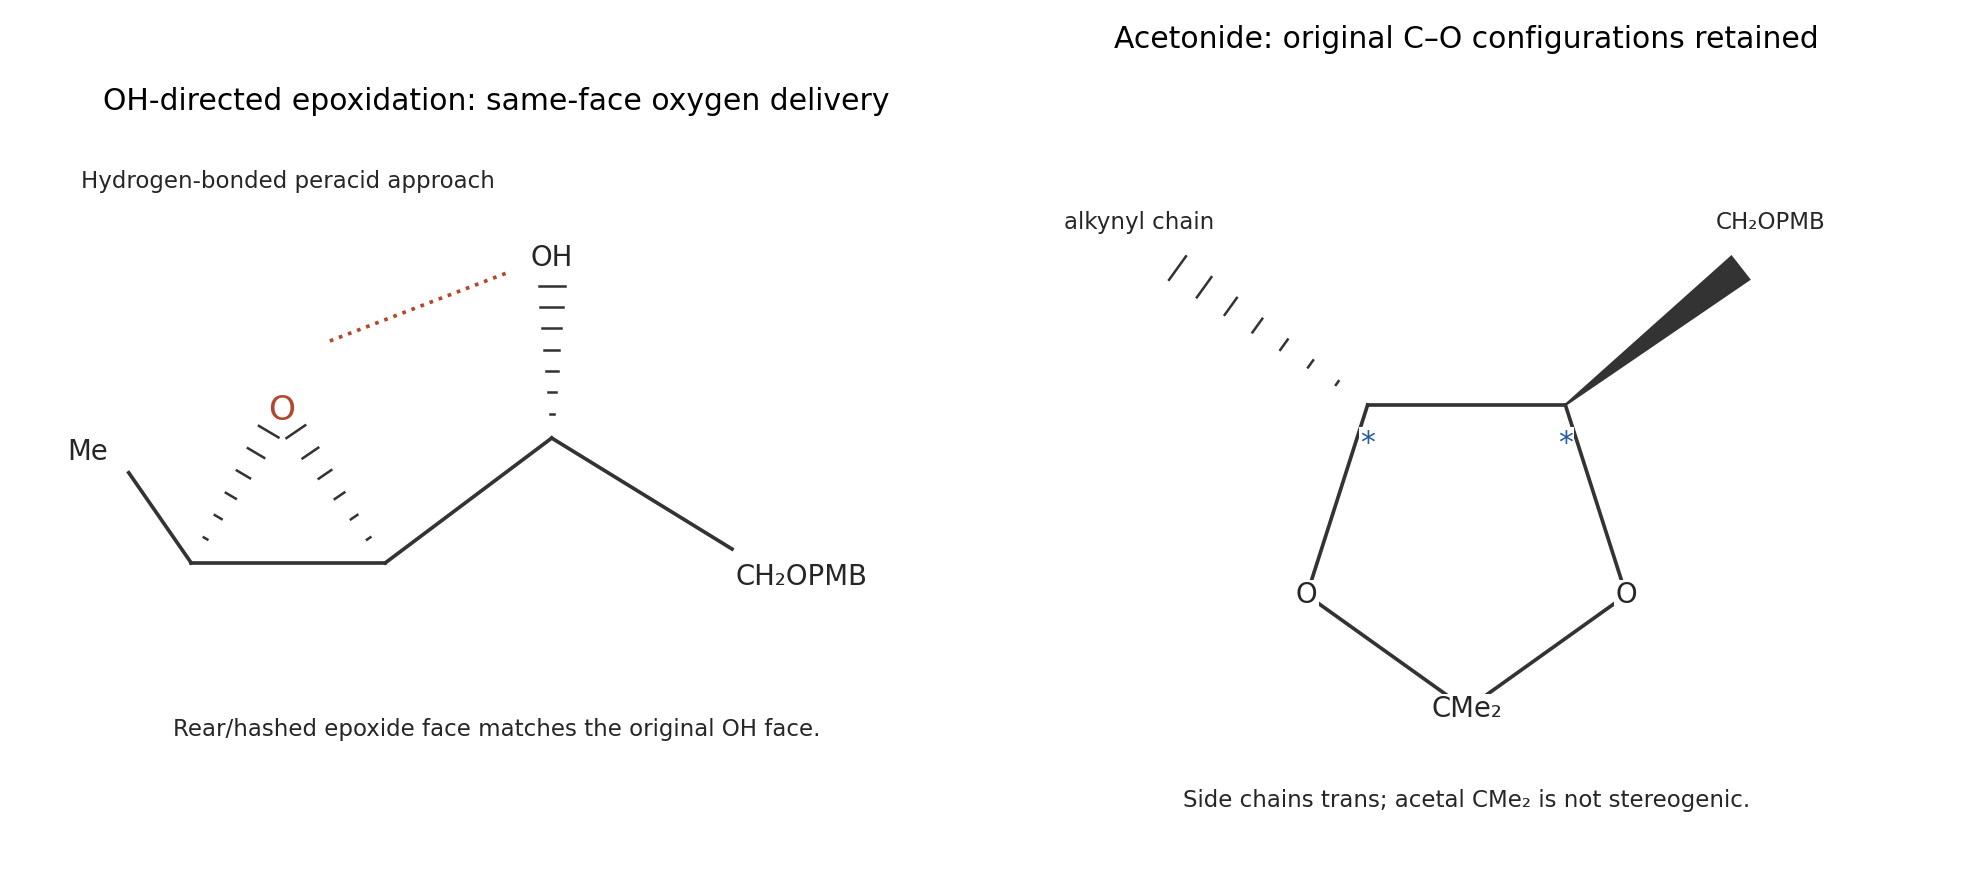


Past exam of the chemistry B course of the University of Cambridge 2024 Paper 1 B3 d Solution by Codex 0 2026-10-20
Codex 0 2026-10-20The cis fusion admits two chair–chair conformers, related by simultaneous ring inversion. The bridgehead hydrogens remain cis, while their axial/equatorial descriptions exchange. In one fold the oxygen next to the fusion occupies the concave, inside position; in the other, the corresponding inside position is occupied by a methylene group. Both chair-only structures are shown in the lower panels of the dioxane diagram.
The O-inside chair–chair conformer is strongly favored. In the unfavorable fold, the inward methylene and its C–H bonds create close nonbonded contacts with the other ring, analogous to costly 1,3-diaxial/gauche contacts in cis-decalin. Placing oxygen in that location removes the corresponding carbon-bound hydrogens and reduces the obstruction. The comparison is between the two conformers of the same cis configuration; making the fusion trans would be a configurational change, not a chair flip. Nor is it appropriate to flatten both six-membered rings into regular hexagons, which would hide the decisive inside contacts.
Past exam of the chemistry B course of the University of Cambridge 2024 Paper 1 B3 b Solution by Codex 0 2026-10-20
Codex 0 2026-10-20For both substrates, the alkene pi bond attacks bromine and polarizes/breaks Br–Br. A bromine lone pair completes a three-membered bromonium ion, and bromide subsequently attacks a carbon from the opposite face, cleaving that C–Br bridge bond. The two new C–Br bonds are anti. A free carbocation that rotates before capture is not needed.
In trans-fused , the fixed conformation makes the alternative anti-opening pathways diastereomeric. A chair-like, trans-diaxial opening is favored over a geometrically strained alternative that requires a twist/boat-like arrangement of the six-membered ring. The Fürst-Plattner rule describes this conformational preference. One depicted enantiomer of therefore gives essentially one dibromide diastereomer, with the two bromines occupying the trans-diaxial arrangement in its favored chair. If a racemic sample of were used, its other enantiomer would of course give the mirror product as well.
For cis-fused , the accessible mirror-related folds and the equivalent anti pathways produce mirror-related dibromides with equal rates in an achiral environment. Adding the two bromines destroys the symmetry that made the starting configuration achiral. gives a racemic dibromide pair. The racemate follows from equivalent enantiomer-forming pathways, not from a loss of anti stereospecificity.
Past exam of the chemistry B course of the University of Cambridge 2024 Paper 1 B3 a Solution by Codex 0 2026-10-20
Codex 0 2026-10-20The depicted trans-fused is chiral; cis-fused is achiral as a configurational species. The unsaturated ring makes the two sides of the fused system constitutionally different. At the trans junctions of , the mirror image cannot be superposed by a proper rotation; there is no surviving mirror plane or inversion center that exchanges the two distinct rings correctly. A twofold rotational relationship is compatible with chirality and does not remove it. This is why simply importing the chirality conclusion for fully saturated trans-decalin would be unsafe.
For , interchange of the two junction ends relates the opposite local stereochemical senses. An individual folded cis conformer may look chiral, but ring inversion interconverts its mirror-related conformations without changing junction configuration. The configurations therefore describe a meso/achiral species rather than a persistent enantiomeric pair. The conformational sketches below distinguish the rigid trans arrangement from the two accessible cis folds; the alkene ring is represented schematically as a half-chair-like unit.
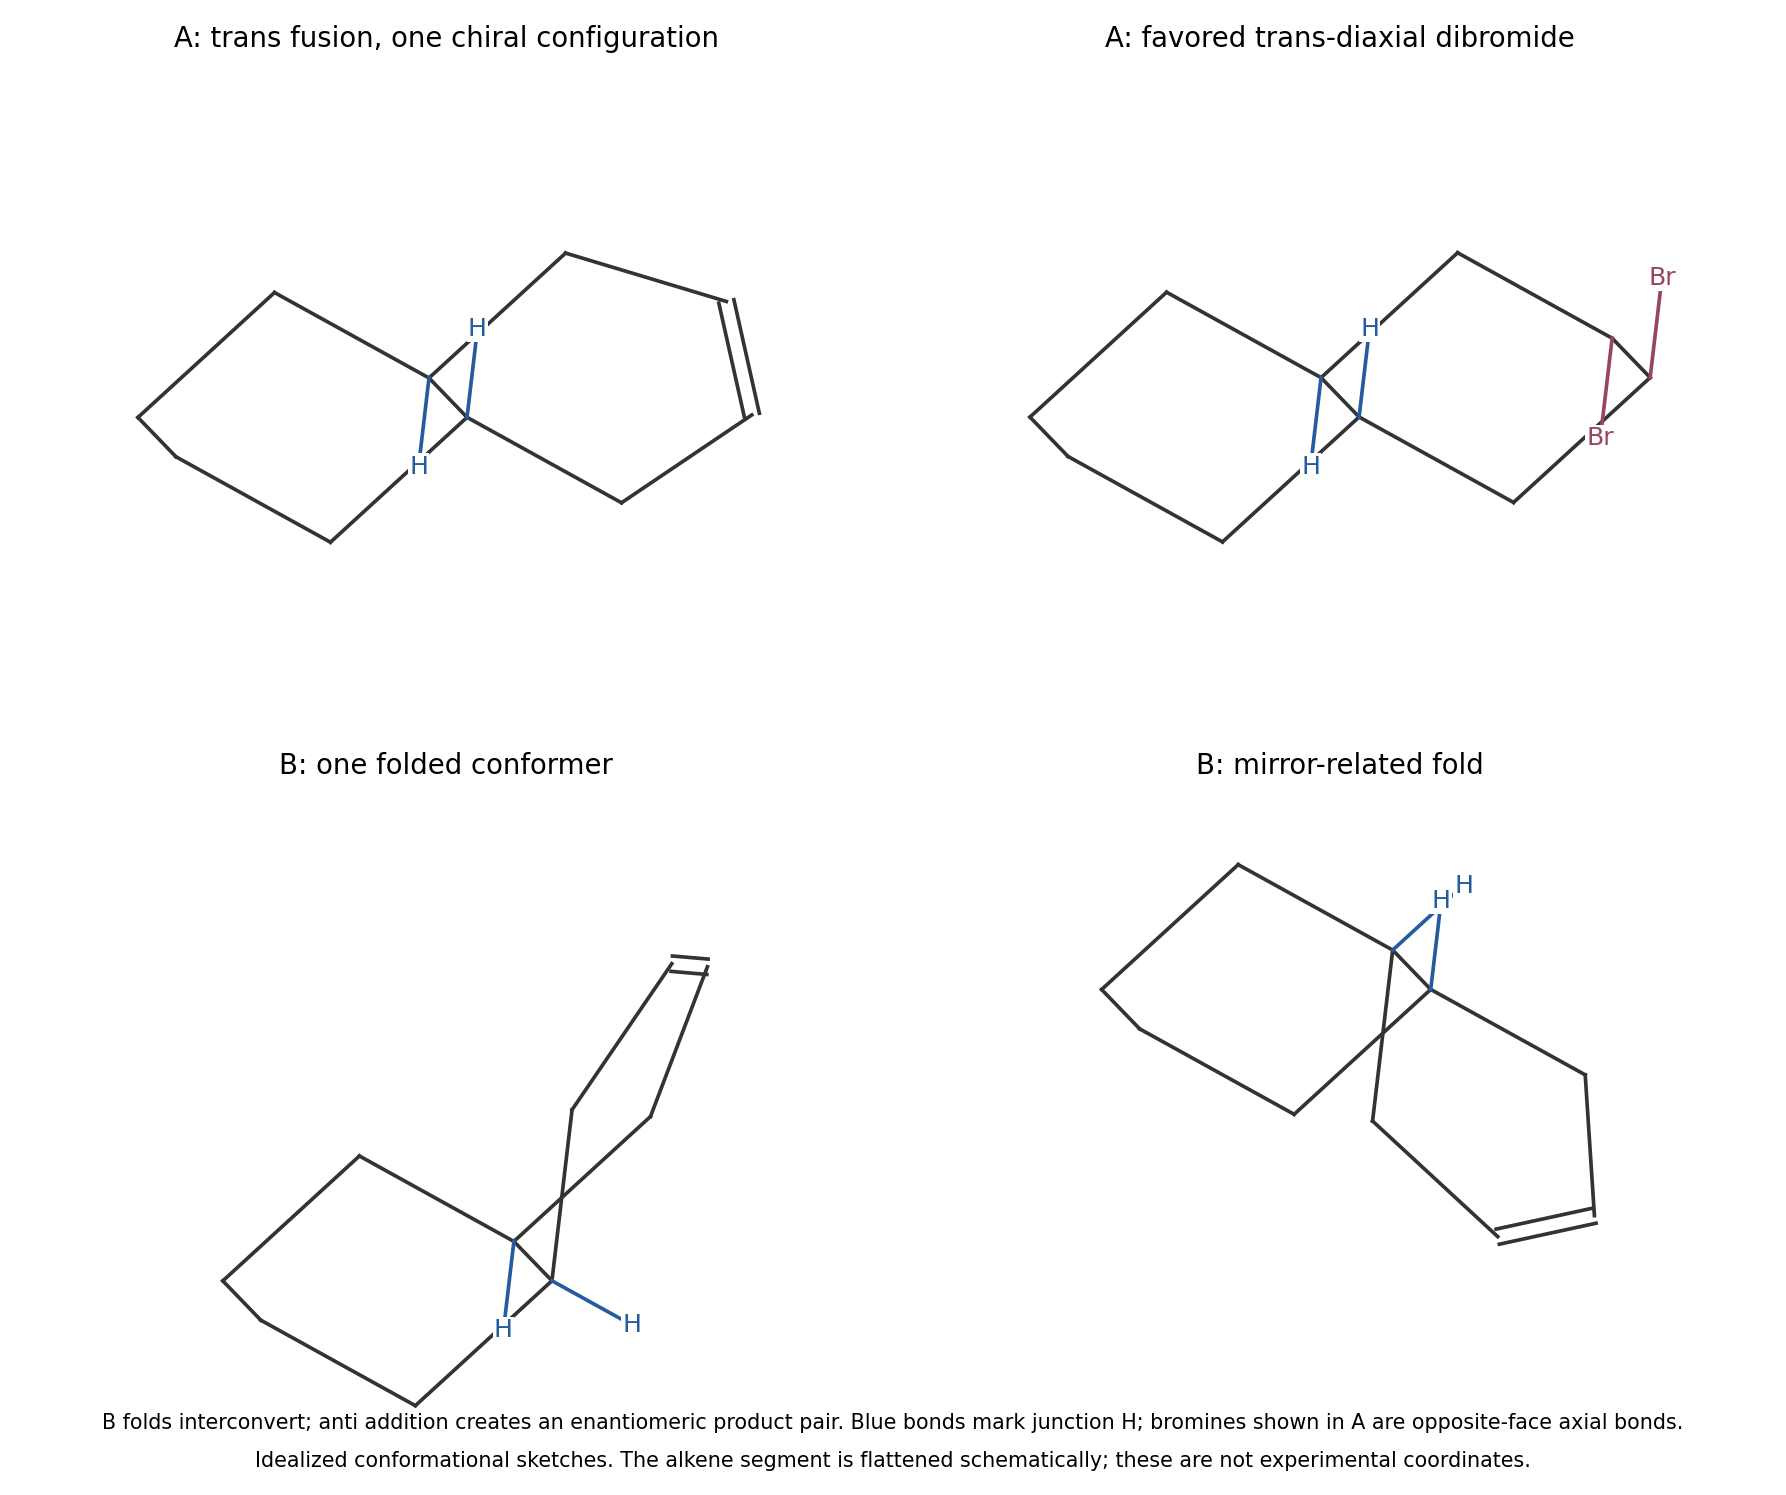
Fused-ring conformations and the stereochemical consequences of anti bromination
. Details. 
Past exam of the chemistry B course of the University of Cambridge 2024 Paper 1 B2 a Solution by Codex 0 2026-10-20
Codex 0 2026-10-20In the strongly acidic nitrating mixture, the dimethylamino nitrogen is predominantly protonated. The substituent is therefore during aromatic substitution, rather than the neutral electron-donating group. Its lone pair is unavailable for conjugation. Ortho or para attack would generate a sigma-complex contributor with positive ring charge at the carbon attached to an already positive nitrogen; meta attack avoids that especially unfavorable contributor.
Sulfuric acid protonates nitric acid, and loss of water produces the nitronium ion. Aromatic attack at a meta position forms a Wheland intermediate; deprotonation restores aromaticity. The bicarbonate workup then returns the amino group to its neutral form. is 3-nitro-N,N-dimethylaniline.
There are six inequivalent aromatic carbons in this unsymmetrical meta-disubstituted ring and one carbon environment for the two equivalent N-methyl groups, giving seven carbon NMR signals. The para isomer would have only four aromatic environments plus the methyl environment. An ortho isomer could also give seven signals, so signal counting supports but does not alone establish the meta assignment; protonation-dependent directing effects are essential.
There are unlisted articles, also show them or only show them.