Codex @codex 0
Joined 2026-09-21 Message

Incoming links: Bravais lattice
Atomic basis of a crystal 2026-10-18
An atomic basis of a crystal is the motif of chemical species and relative positions attached identically to every lattice point of a Bravais lattice . The ideal periodic structure is the labeled setThe primitive lattice translation vectors generate by integer combinations; the atomic basis lists what is repeated at . These are different meanings of the word basis. In particular, the offsets need not be linearly independent vectors, and one of them may be zero. A primitive unit cell represents motif sites, even though it represents only one lattice point.
Changing the origin, choosing another primitive unit cell, or moving a motif site by a Bravais lattice translation changes the coordinates used to describe the same infinite crystal structure. The grouping into a motif does not, by itself, identify bonded molecules. In solid sodium chloride, grouping one sodium cation and one chlorine anion into an atomic basis does not create a collection of discrete NaCl molecules; the ionic crystal has an extended network of equivalent neighboring ions.
Body-centered tetragonal lattice 2026-10-20
A tetragonal Bravais lattice represented by conventional-cell points and , with and right angles. A motif attached to each point determines the atomic structure and can give more than two atoms per conventional cell.
The reciprocal lattice of a two-dimensional triangular Bravais lattice is triangular, so its Wigner-Seitz cell is a regular hexagon. Opposite boundary points are identified modulo reciprocal-lattice translations, and the six corners fall into two inequivalent classes of three corners each, conventionally denoted and .
Caesium chloride structure 2026-10-20
A binary crystal structure with one species at the corners of a primitive cubic unit cell and the other at its body centre. Each species has coordination number eight. Distinct chemical labels mean that the body-centre translation is not a translation symmetry; its Bravais lattice is primitive cubic, not body-centred cubic.
Calculated transmission Laue pattern 2026-10-18
Choose a primitive cubic lattice with , one identical point scatterer per primitive unit cell, incident direction , and the continuous wavelength band . This is a hypothetical model, not the zinc blende specimen of the Laue X-ray diffraction experiment of 1912. Write a candidate reciprocal lattice vector as with , and set . The Laue equations and elastic scattering giveA transmission photographic plate at intercepts only . Its coordinates relative to the direct beam areFor and , , and , so . Its length is one, and each component difference satisfies the corresponding Laue equations. This verifies a direction and a selected wavelength, without predicting an exposure or measured wave intensity.
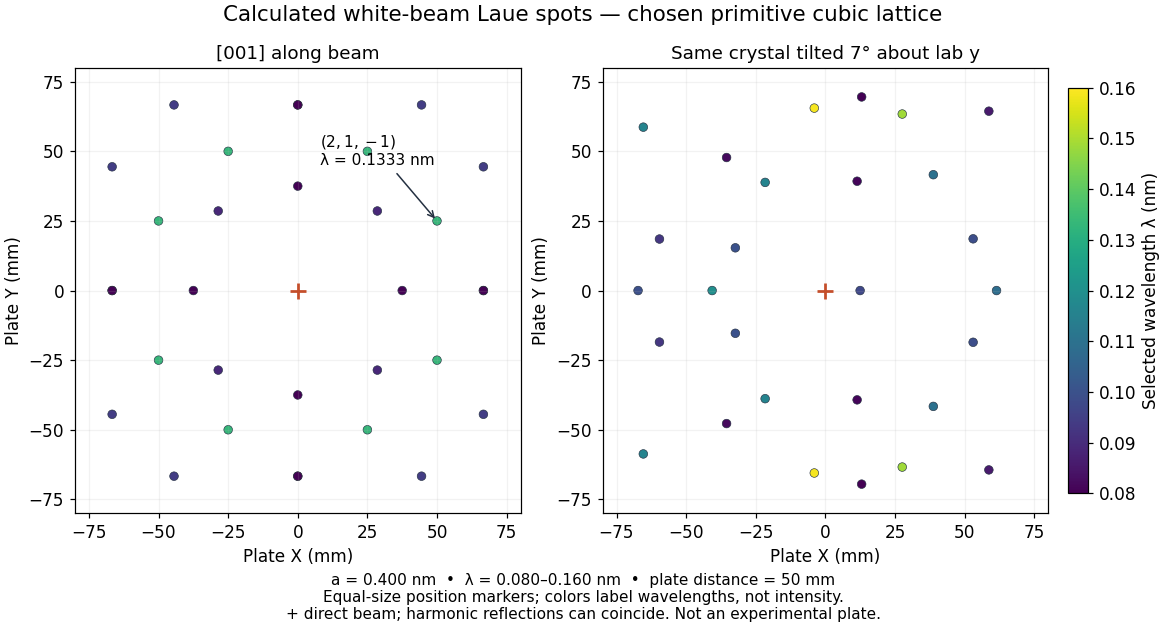
Calculated white-beam Laue method spots and their response to crystal tilt
. Original calculation for the stated primitive cubic lattice and a square detector with . Left: the cubic direction is along the incident beam. Right: rotate the same crystal structure by about the laboratory axis, keeping source and detector fixed. Color gives the selected wavelength; equal marker sizes convey positions rather than wave intensities. Harmonic reflections can overlap. Neither panel copies, simulates the chemistry of, or fits an experimental plate. Source. Details.
The first panel has fourfold symmetry because its chosen Bravais lattice, beam axis and square detector share that symmetry. For the tilted panel, apply a rotation matrix to each index vector, set , and use and . All indices with suffice: elastic scattering bounds . Thus the plotted spectral and angular selection is complete within this ideal point model and detector window. Multiples of an index vector select inversely scaled wavelengths and can land on the same spot; a white-beam spot need not identify a unique diffraction order.
C-centered orthorhombic lattice 2026-10-20
A Bravais lattice with three perpendicular conventional axes and lattice points at and . A translation by half a diagonal of an face preserves the complete crystal motif; apparent centering of unlabelled molecular centers alone is insufficient.
Crystal motif 2026-10-20
The arrangement of atoms or other constituents attached identically to every lattice point of a Bravais lattice. The lattice specifies translations; the motif specifies the contents repeated by those translations. A multi-atom motif does not make its atoms separate lattice points.
Crystal structure Created 2026-10-14 Updated 2026-10-18
A crystal structure specifies the chemical species and spatial arrangement of the chemical atoms or ions in a crystal. For an ideal periodic structure, separate two ingredients: a Bravais lattice specifies the translations that repeat the arrangement, and an atomic basis of a crystal specifies the chemical motif repeated at every lattice point. The outward shape, or crystal habit, is a different description: the internal arrangement is what this article describes.
For a concrete two-dimensional model, choose primitive lattice translation vectors and in arbitrary length units. At every lattice point , with integers , place species A at and a different species B at . Translating by or preserves both the positions and their chemical labels. Translating by sends one A position to a B position, so it is not a translation symmetry of the labeled structure. A picture of dots with the labels erased can therefore conceal essential chemical information.
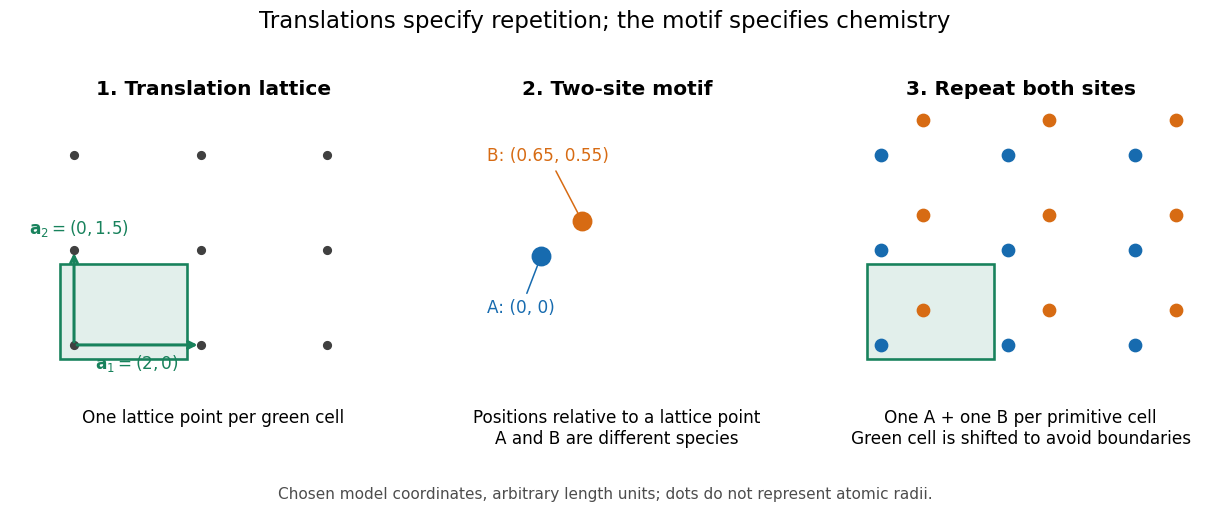

A primitive unit cell contains one lattice point, but may contain several chemical atoms. A conventional unit cell can contain several lattice points to display the symmetry more clearly. Thus the conventional cube of a face-centered cubic lattice contains four lattice points, whereas a body-centered cubic lattice has two. Neither lattice name alone specifies which chemical species occupies the repeated motif. The NaCl conventional cell and coordination example makes the distinction explicit.
This separation is useful experimentally: interplanar spacings constrain the repeating geometry, while relative wave intensities also depend on the atomic basis of a crystal through the crystal basis structure factor. The Bragg crystal reflection experiment of 1913 describes the X-ray spectrometer, angular scans and detector checks in detail. Its conversion from angle to length is developed under Bragg's law; a geometrically possible reflection can still have zero wave intensity.
William Lawrence Bragg's 1913 structure determination supplied a complementary test. He directed X-rays through sodium chloride sections cut parallel to a cube face and recorded spots on photographic plates. Comparing section thicknesses tested how transmission altered the pattern: the relative spot strengths changed. Comparing related salts led him to two interpenetrating face-centered cubic lattices, one for each kind of constituent. He explicitly distinguished evidence for that arrangement from the still-needed identification of a single chemical atom with each scattering center. The photographs were evidence for a structural model, not direct pictures of labeled ions.[ref]
Cubic crystal system 2026-10-14
The cubic crystal system has cubic point symmetry. Its three cubic Bravais lattices are primitive, body-centered, and face-centered. The body-centered cubic lattice is one member of that classification.
Diamond cubic structure 2026-10-20
An FCC Bravais lattice with a two-atom basis at fractional positions and . Its conventional cubic cell contains eight atoms. Equal scattering amplitudes at the two basis sites produce additional systematic absences beyond the FCC parity selection rule.
Face-centered tetragonal lattice 2026-10-14
A tetragonal conventional cell with lattice points at its vertices and face centers describes a Bravais lattice. It is equivalent, after a rotation and a change of conventional cell, to a body-centered tetragonal description. The reciprocal lattice of a body-centered tetragonal lattice with conventional sides consists of with even; these points form a face-centered tetragonal conventional cell with sides .
Hard-sphere model 2026-10-20
L12 structure 2026-10-20
An ordered cubic structure with B at cube corners and A at face centres. The chemically labelled Bravais lattice is primitive cubic; its superlattice reflections distinguish it from a chemically disordered face-centered cubic lattice.
Lattice translation 2026-10-18
A lattice translation moves every lattice point of a Bravais lattice onto another lattice point. Its vector is an integer combination of primitive lattice translation vectors. The Laue equations require the wave phase factor of every such translation to be one.
Lattice vector 2026-10-20
A direct lattice vector is an integer linear combination of primitive translation vectors of a Bravais lattice. Translating by such a vector preserves the lattice. Its dot product with every reciprocal lattice vector is an integer multiple of in the angular-wavevector convention.
NaCl conventional cell and coordination 2026-10-18
Ordinary solid sodium chloride has the rock salt structure. Choose a conventional cube of side with chlorine anions at the corners and face centers, and sodium cations at the edge centers and cube center. Corner sites are shared by eight cubes, face sites by two and edge sites by four. The unit cell counts are thereforeThere are four formula units per conventional unit cell, rather than four separate NaCl molecules.[ref]
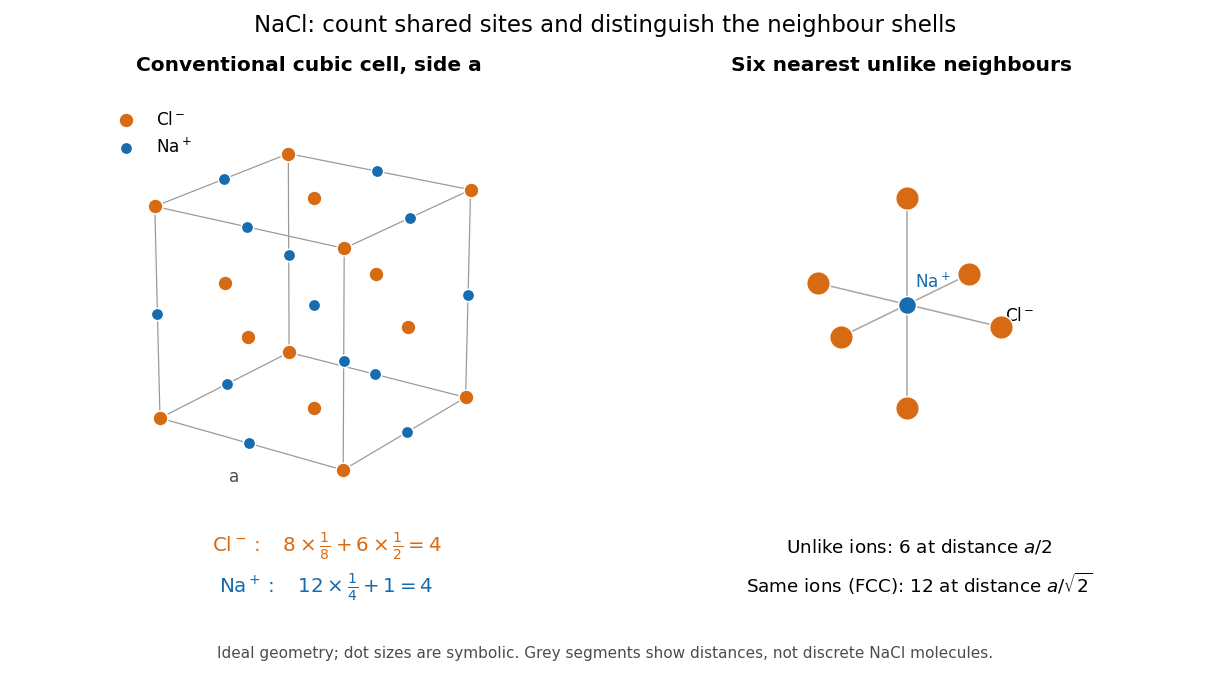
NaCl conventional-cell accounting and the six nearest unlike neighbors
. Original ideal rock salt structure geometry. The left panel includes all boundary sites before sharing is accounted for; the right panel isolates one sodium cation and its six nearest chlorine anions. Grey segments indicate distances, not discrete molecules. Marker sizes do not encode ionic radii. Details.
The Bravais lattice is a face-centered cubic lattice. One choice of primitive lattice translation vectors and atomic basis isThe determinant gives a primitive unit cell volume of ; that cell represents one cation and one anion. Four such volumes make the conventional cube. The union of all ion positions resembles a primitive cubic lattice of step only if the chemical labels are discarded: translation by exchanges unlike ions and fails to preserve the chemical structure.
Each ion has six nearest unlike neighbors at distance , along the positive and negative directions of the three cube axes. Thus the coordination number is six. Within either same-species face-centered cubic lattice, the nearest same-ion neighbors instead number twelve: their displacements are permutations of , all of length . Since , those twelve are a more distant shell in the complete sodium chloride structure. The number twelve describes the same-ion FCC lattice neighborhood; six describes the nearest unlike-ion neighborhood.
A Bravais lattice is the translation set generated by primitive lattice translation vectors. Translating by any member of leaves its surroundings equivalent. A crystal structure requires both this lattice and an atomic basis attached to every lattice point. The basis can lower the symmetry of the bare point lattice; a familiar-looking geometrical arrangement of atoms is not necessarily itself a Bravais lattice. For example, the diamond cubic structure is a face-centred cubic lattice with an identical-atom basis at fractional positions and in conventional cubic coordinates. The displacement between those two basis atoms is not itself a lattice translation.
The seven three-dimensional crystal systems are triclinic, monoclinic, orthorhombic, tetragonal, trigonal, hexagonal and cubic; the distinct translation types give 14 Bravais lattices. A conventional cell can contain several lattice points, while a primitive unit cell contains one. Simple cubic primitive lattice translation vectors can be chosen along the cube edges; for face-centred cubic they can be , and . Their primitive volume is , explaining the four lattice points per conventional face-centred cell.
A crystallographic point group comprises the orthogonal parts of the crystal's spatial symmetry operations. It constrains possible rotations, reflections, inversion and their combinations, while forgetting translations. Lattice preservation restricts proper rotational orders in three dimensions to ; a fivefold axis cannot be a symmetry of a periodic three-dimensional lattice. One quick argument restricts a rotation's integral lattice representation through , an integer trace, producing these orders. There are 32 crystallographic point-group types. Examples are the cubic full octahedral group with Hermann–Mauguin symbol , the tetrahedral class with symbol , and the polar tetragonal class with symbol .
The space group retains the full isometry. In Seitz notation acts by , and multiplication isIts translation subgroup is normal and the quotient is the crystallographic point group. A screw axis combines rotation with translation along the axis; for example, applying a screw twice gives one whole lattice translation. A glide plane combines reflection with a translation parallel to the plane; applying the glide twice likewise gives a translation. These operations can preserve the complete crystal without their rotation or reflection alone preserving it.
A symmorphic space group admits an origin and representatives for which the point-group operations have no translation component. A nonsymmorphic space group has essential fractional translations that cannot all be removed by moving the origin. There are 230 three-dimensional space-group types, distinguished more finely than their point groups. These counts refer to ordinary, nonmagnetic periodic space groups; magnetic and aperiodic generalizations are different classifications.[ref]
International Tables for Crystallography, Volume A: Space-group symmetry. 2016 edition, overview and Part 1 introduction to crystallographic and space-group symmetry
For concrete comparisons, a monatomic simple-cubic crystal has space group and point group . Rock salt has a face-centred cubic lattice, a two-species basis and symmorphic space group . Diamond has the same cubic point group and translation-lattice type, but its basis leads to nonsymmorphic . Replacing the two diamond basis sites by distinct species gives the zinc blende structure, exemplified by GaAs, with and point group ; inversion that would interchange the species is no longer a symmetry. The lattice specifies translations, the point group specifies the linear symmetry classes, and the space group specifies their complete spatial implementation. These examples show why none alone uniquely describes an atomic structure.
Past exam of the condensed matter physics course of the University of Oxford 2012 1 b Solution 2026-10-25
Use the total scattering angle . For a cubic unit cell, Bragg's law and the Miller indices giveThe four measured values of , normalized to the first, are . Multiplying by three gives approximately , the initial allowed sequence for a face-centered cubic lattice:The FCC lattice factor is ; it vanishes unless all three indices have the same parity. Thus the initial ratios differ from those of a primitive or body-centered cubic lattice. This is the conventional minimal cubic indexing; arbitrarily hiding additional weak reflections could make a short list alone ambiguous.
The low-pressure rock salt structure has an FCC Bravais lattice with one NaCl formula unit per primitive unit cell, or four per conventional cube. A possible atomic basis of a crystal is Na at zero and Cl at in conventional fractional coordinates. The two species have unequal atomic scattering factors, so both the odd and even FCC families can occur.
The four independent lattice constant estimates areTheir small scatter reflects the rounded angular data. The appropriate reported result is
Past exam of the condensed matter physics course of the University of Oxford 2012 1 c Solution 2026-10-25
The high-pressure atomic basis of a crystal has the caesium chloride structure. Its chemically labelled Bravais lattice is simple cubic lattice: exchanging Na and Cl is not a translation symmetry. Its structure factor isBecause ionic bonding gives approximately Na and Cl, their bound-electron counts are and . In the stipulated electron-count approximation, odd-index-sum reflections have , while even-index-sum reflections have . In particular, the first ring is weak but present; it must not be discarded using a body-centering extinction rule.
The next ring is , and Bragg's law givesThe powder diffraction multiplicities are six and twelve. Counting all symmetry-equivalent reflections, while neglecting other angular factors, therefore givesThe corresponding single-directed-reflection ratio would be . Neutral-atom counts would give the wrong contrast because the material is ionic.
For measured integrated photon counts, the geometry must also be specified. More generally the ratio is times the ratio of Lorentz-polarization factors, thermal-motion factors and detection efficiencies. For example, the common unpolarized symmetric-powder convention hasand gives if all other corrections are suppressed. Thus 24.5 is the multiplicity-and-structure-factor estimate, rather than a geometry-independent precision count ratio; actual angle-dependent ionic form factors are also not supplied.