and . Both metals are bridged by the two diisopropylamide nitrogen atoms in a four-membered Li–N–K–N ring. The smaller lithium ion accommodates the two amides plus one THF oxygen, while the larger potassium ion has space for two THF donors and more readily adopts the higher coordination number. The principal Li–N and Li–O contacts should be shorter than their potassium counterparts because of the much smaller lithium ionic radius. A larger number of donors does not imply shorter individual K–donor bonds.
Protonolysis of both magnesium–butyl bonds givesThus A is magnesium bis(hexamethyldisilazide), and the gas is butane. Addition of the corresponding lithium amide gives B = , a lithium tris(amido)magnesiate.
A useful connectivity sketch is with : magnesium binds three approximately coplanar nitrogen atoms, two of which also bridge to lithium. The remaining nitrogen is terminal at magnesium. Lithium can gain additional intramolecular contacts to methyl groups on the silylamide substituents. These contacts and the steric envelope of the bulky ligands help satisfy coordination without intermolecular aggregation. “Monomeric” here means one LiMg formula unit, not a single metal atom or an isolated bare counterion.
C and the dimer of D have the same composition, , but different metal ordering around an eight-membered metal–nitrogen ring. C has adjacent Cu sites and adjacent Li sites; in the conventional Gilman dimer D, Cu and Li alternate around the four metal positions. D is represented by per formula unit. The salt by-product is LiCl.
A TMP anion has formula and mass about 140.25. Therefore the mass per empirical unit is , giving for the dimer, consistent with . A monomer would be near 351 and is excluded. In C, inequivalent Cu/Cu, Cu/Li and Li/Li amide surroundings, combined with the constrained TMP orientations, permit the three methyl environments in the observed 1:2:1 integral ratio. D has the higher exchange-averaged methyl equivalence needed for one methyl signal. One signal need not imply that all instantaneous solid-state methyl environments are identical.
The remaining formulae follow from atom and charge balance:E can itself aggregate; its formula here specifies the copper-amide unit. F is a mixed organyl/amido Gilman cuprate, and G combines one F unit with one D unit. The distinct metal-ordering isomers and these mixed aggregates have experimental precedent.[ref]
A new route for the efficient metalation of unfunctionalized aromatics by Andrew J. Peel and others. 2019, metal-exchange isomers and mixed organyl/amido aggregates
F is an eight-membered ring with sequence , closed back to the first Cu. Each N belongs to TMP; each ipso carbon belongs to a phenyl group. Each Cu is approximately linear between one TMP nitrogen and one phenyl ipso carbon. Each Li contacts a TMP nitrogen and the phenyl group bound to the other Cu, so the dimer is assembled head-to-tail rather than from two independent separated ion pairs.
G has the eight-membered sequence , closed to Cu. The three N atoms belong to three TMP groups and the carbon belongs to the single phenyl group. Both copper sites remain approximately two-coordinate. The Li next to phenyl can also interact with its aromatic surface; this secondary contact need not have the same hapticity as in F. The figure gives a connectivity projection, not idealized square bond angles or an X-ray metric reconstruction.
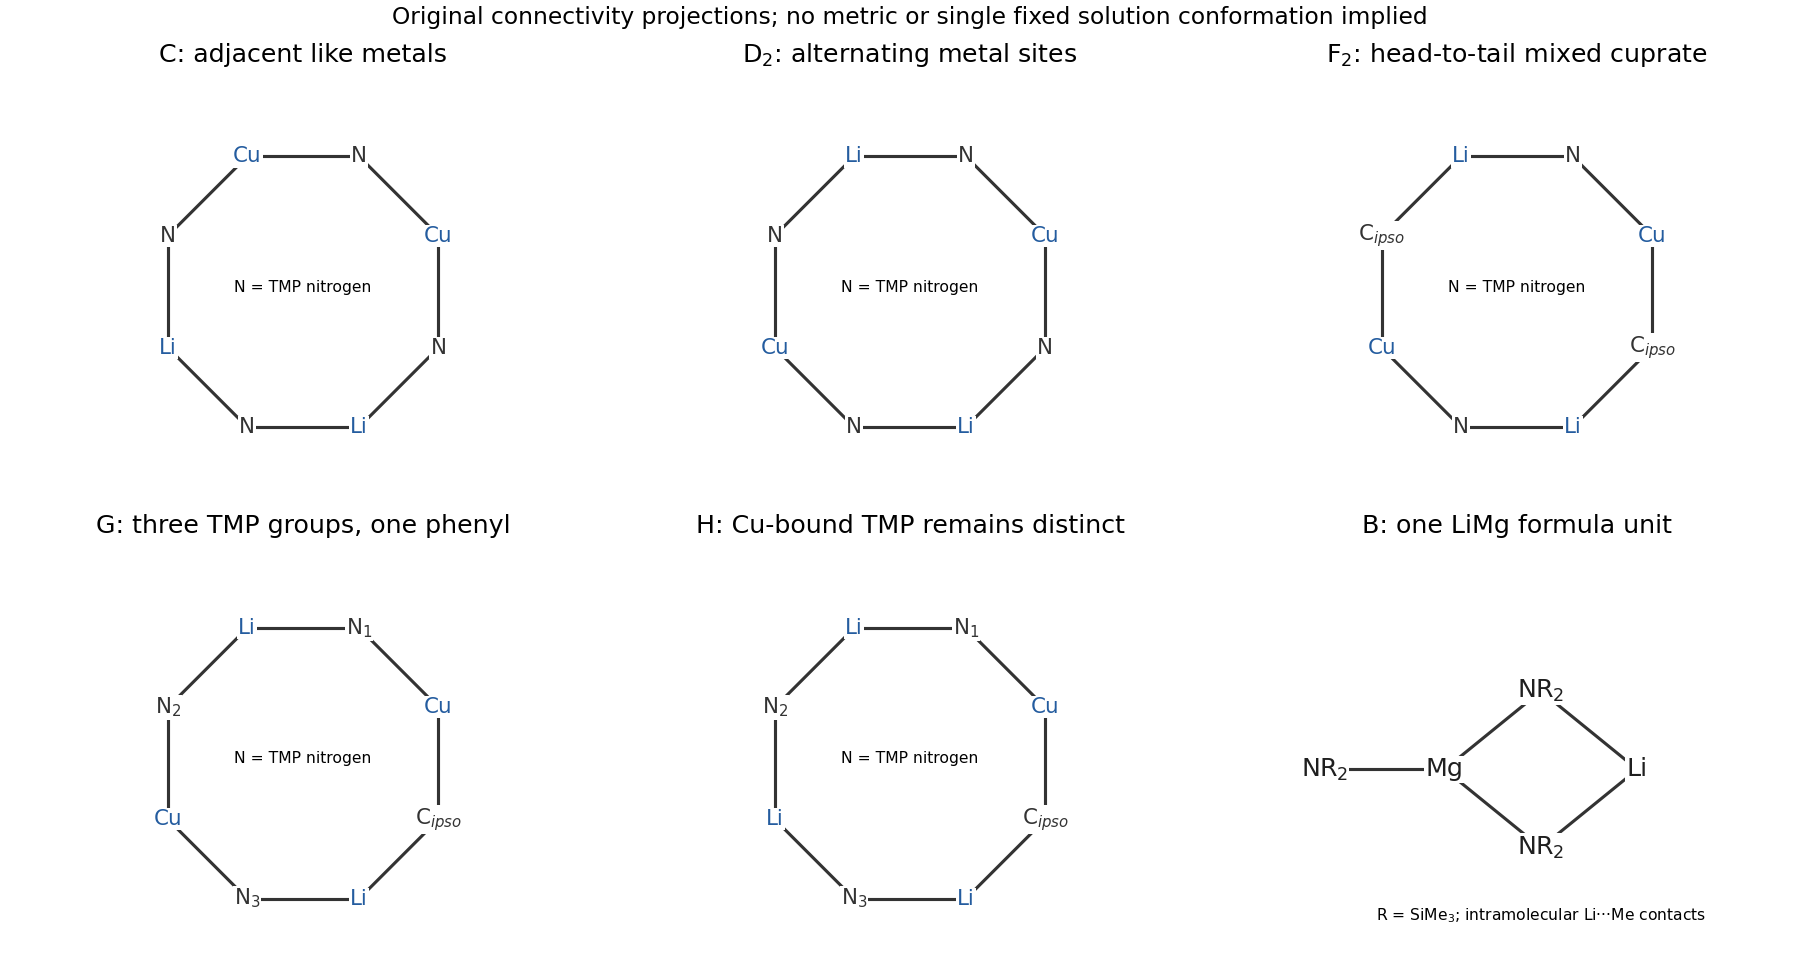
Original connectivity diagrams for amido-metal aggregates and the exchangeable lithium-amide segment
. Details. 
Stoichiometry gives H = , formally F plus two LiTMP units. A compatible ring connectivity is , closed through a lithium–phenyl interaction. The TMP attached to copper, N, remains chemically distinct; N and N belong to the more labile lithium-amide part.
Reversible detachment and reattachment of a symmetric segment can exchange its two amides and reassign which Li occupies the phenyl-contacting or nitrogen-bridging positions. It can therefore exchange all three Li sites and the two Li-associated TMP groups while retaining the Cu-bound TMP group. Fast TMP-ring conformational inversion can further average methyl environments without exchanging entire ligands. These are different kinds of motion; simple ring inversion alone cannot explain exchange of the lithium sites. Such a dissociative interpretation is consistent with the experimentally studied lithium-rich aggregate, but the exchange pattern does not determine a unique transition state or prove that every bond breaks in one concerted event.
Each fragment leaves two electrons after allocating one aluminium valence electron to its monovalent cyclopentadienyl ligand. Four fragments therefore give eight cluster electrons. In the tetrahedral arrangement, fragment lone-pair/donor and vacant acceptor orbitals combine into delocalized cluster orbitals. The occupied low-energy set has symmetry, analogous to a filled 1S and 1P shell; higher combinations remain empty. This is a multicentre description, not six independent two-electron Al–Al bonds, which would require twelve electrons.
A schematic molecular-orbital diagram should show the occupied nondegenerate level below the occupied triply degenerate set, separated from the empty higher levels. Detailed energies and ordering of the empty levels require an orbital calculation. On descending Group 13, stabilization of the pair and reduced favourable – mixing can weaken the tendency towards the same compact Al-like bonding pattern; metal–metal distances and preferred aggregation change. Ligand size, donor/acceptor properties and hapticity also tune fragment orbital energies and steric access, so neither substitution of the metal nor of Cp* is electronically innocent.
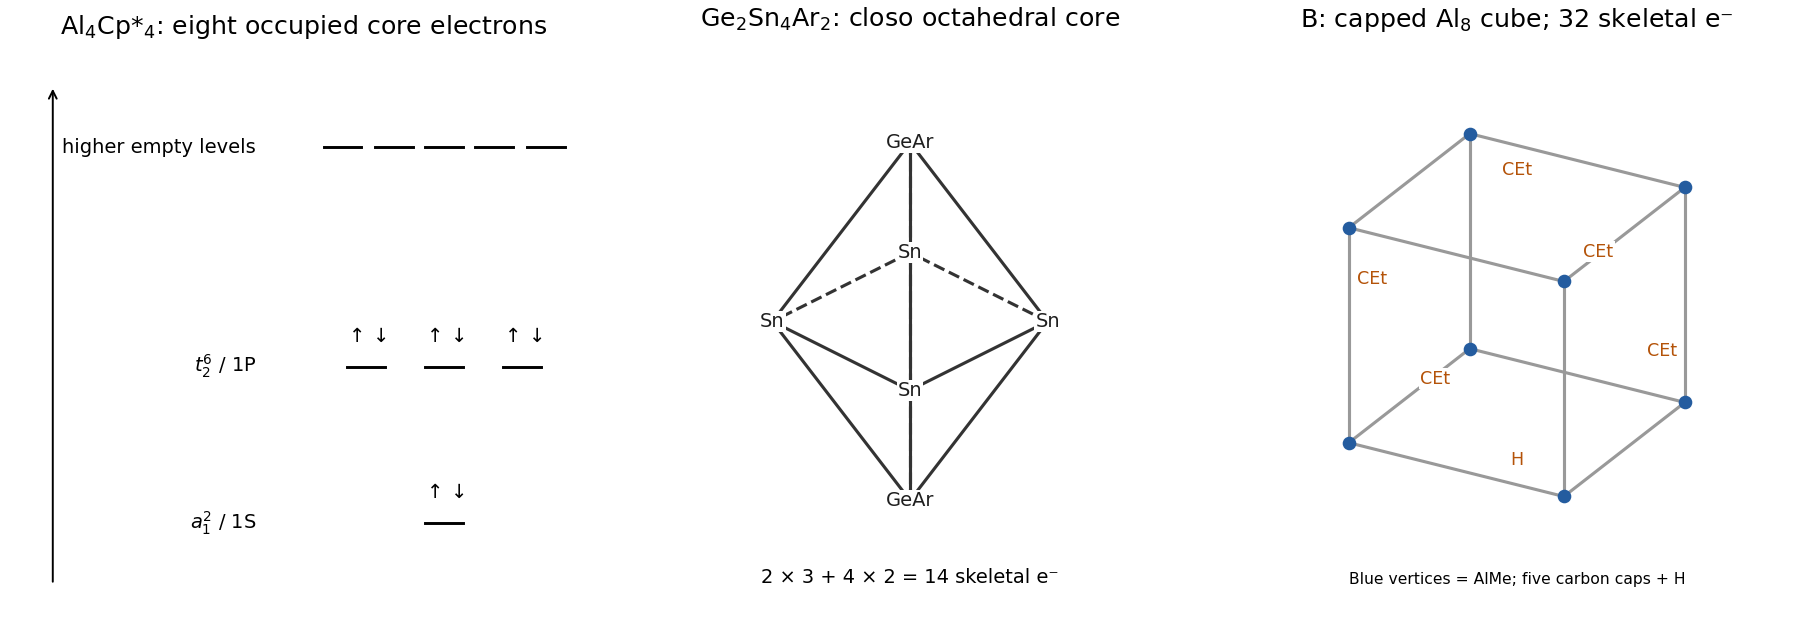
Original cluster orbital filling, octahedral electron counting and capped-cube connectivity
. Details. 
A shell with angular momentum holds electrons. Successively filling the shells listed in the source gives the closed-shell totalsFor the delocalized count is , matching . The tetrahedral aluminium core is consequently consistent with the jellium model. Counting the same eight electrons as though the tetrahedron were an ordinary closo Wade cluster would fail: a four-vertex closo framework would require skeletal electrons. The competing models use different physical assumptions and should not be interchanged just because both involve a polyhedron.
In a skeletal description, each of the four naked Sn atoms retains an outward lone pair and contributes two electrons to the framework. Each aryl-substituted Ge atom contributes three electrons after assigning one to its Ge–aryl bond; it is not assigned an additional spectator lone pair in this counting. ThusThe PSEP prediction is a six-vertex closo octahedron, with the two protected Ge vertices and four naked Sn vertices; distortion does not change the topological classification. The alternative gross delocalized count is not a listed jellium shell closure. Subtracting a lone pair from every atom would incorrectly remove electrons from the substituted Ge framework and give the wrong classification.
The central silicon contributes four electrons to the delocalized metal-core count. Six monovalent Cp* ligands each withdraw one electron, givingThis is a jellium closed shell, through the filled 2P shell. The label identifies an encapsulated silicon with eight aluminium contacts; it should not be counted as an ordinary exterior Wade vertex with an outward lone pair. The gross electron count rationalizes shell closure but does not alone reconstruct every Al–Al bond or all ligand positions.
The revised question page explicitly gives the cluster as . Its charge supplies two electrons, soa jellium shell closure, through 1G. The reported structure has a Ga atom inside an eleven-atom Ga shell, with ten further ligand-bearing Ga units outside that core.[ref]
A Metalloid Cluster Compound with a Variation of the Ga22 Framework by Andreas Schnepf and others. 2002, central Ga@Ga11 core and ten external GaR units
The canonical exam copy omits the 2− charge. Taken literally as a neutral formula it gives 56 electrons, which is not a listed shell closure; assigning 58 without identifying this difference would be unjustified. The revised public source resolves the discrepancy rather than requiring an invented charge.[ref]
Revised question 33 of the 2022 Cambridge Part III Chemistry Paper 2. physical page 81, part b(iv)
For the neutral aluminium cluster,This closes the listed sequence through 1I and supports the jellium description of a large metalloid cluster. Cp* ligands are counted as monovalent electron-withdrawing surface groups; counting each as a five-electron donor to the entire delocalized core would mix a transition-metal local electron-counting convention with the present shell model. Electron closure is a useful stability criterion, not by itself proof of a perfectly spherical nuclear geometry.
A is the Ru triangle capped by -AlEt, with three edge-bridging hydrides and one Cp* ligand at each Ru: . The 45-proton singlet accounts for three equivalent Cp* ligands, the three-proton upfield singlet for the equivalent bridging hydrides, and the two-proton quartet plus three-proton triplet for one ethyl group.
The revised source has two face-bridging hydrides in the precursor. Substituting the AlEt cap then balances asThus the gas is ethane. Multicentre Ru–H–Ru and Ru–Al bonding stabilizes the capped cluster; describing the AlEt unit as a face-bound low-valent aluminium fragment is more informative than drawing three unrelated localized donor bonds. The canonical precursor formula lacks the second face hydride and would not balance to this product plus two ethanes. The revised formula explicitly repairs that discrepancy.[ref]
Revised question 33 of the 2022 Cambridge Part III Chemistry Paper 2. physical page 81, part c
Eight AlMe vertices form a cube, five faces are capped by -C(Et) units, and the sixth face is capped by a bridging hydride. Choosing the hydride face as the bottom gives four equivalent side carbon caps and one distinct opposite carbon cap. Their ethyl groups give the 4:1 integrations: four methyl groups contribute 12 H and their methylenes 8 H; the remaining ethyl gives 3 H and 2 H.
The eight aluminium-bound methyl groups divide into two sets of four: the four aluminium atoms bordering the hydride face and the four on the opposite face. Each set gives a 12-proton singlet. The single bridging hydride gives the remaining one-proton signal. This matches the complete integration pattern without postulating five inequivalent ethyl caps. It specifies the symmetry-compatible connectivity; NMR alone does not measure every bond length or rule out solution averaging of small distortions.
Use the neutral-fragment skeletal count: each AlMe supplies two electrons, each CEt supplies three and the bridging H supplies one. HenceThere are 13 nonhydrogen skeletal vertices, eight Al plus five C, and pairs. In the usual PSEP classification this is therefore arachno, corresponding to a 15-vertex closo parent with two vertices absent. The μ-H cap contributes an electron but is not counted as a carbon-like skeletal vertex. The aluminium cube is a convenient subframework; its eight corners alone are not the complete Wade skeleton. Calculations on capped cubic carbaalanes likewise identify 16 skeletal pairs for the five-carbon-cap/one-hydride-cap case.[ref]
Electron Counting in Carbaalane Clusters with Cubic Aluminum Core by Benoit Creton and others. 2007, five conical carbon caps and one hydride cap
The requested equality is electrons if each unsaturated unit is counted as a four-electron donor to the chosen extended cluster framework. It needs that additional convention; a count confined to the localized skeleton gives instead.
For , an fragment contributes two skeletal electrons, a singly substituted cap contributes three, and the bridging hydrogen contributes one:For the second formula, let be the contribution assigned to each unsaturated unit. The comparison isThus requires . A bookkeeping partition that includes the two alkene pairs in the extended cluster electron pool givesIn contrast, assigning the double bond entirely to the exoskeletal substituent leaves each corresponding cage carbon with two electrons for the cage:The difference is exactly the two pairs, four electrons, rather than an arithmetic discrepancy. These are different electron partitions: do not compare from one partition with from the other as though both were the same skeletal count, and do not infer a structural classification merely by inserting the extended count into Wade's rules for twelve localized skeletal vertices.
Both the original and revised question pages print the same neutral molecular formula; the revised copy does not supply a charge, a drawing, or an explicit instruction about these electrons.[ref]
Revised question 33 of the 2022 Cambridge Part III Chemistry Paper 2. Part (d)(iii), physical page 81 of the later public bundle; the formula is unchanged from the original question
The primary study identifies the open cluster with the two genuine groups and reports their hydroalumination to give the capped aluminium cube. Its accessible abstract does not establish the orbital partition needed to select the four-electron convention.[ref]New Carbaalanes Containing Clusters of Aluminum and Carbon Atoms by Werner Uhl and others. Abstract: open cluster 2 and its conversion to compound 4; Organometallics 19 (2000), 4536–4543
Conditional conclusion: the advertised -electron equality follows under the explicit extended-framework convention above. Without evidence that those pairs belong in the cluster electron pool, retain the -electron localized count and this qualification. The source checks do not justify asserting a typo or inventing a charge to force agreement.
Articles by others on the same topic
There are currently no matching articles.