
Polymerization converts a monomer, or a mixture of monomers, into a polymer: repeated chemical reactions establish a connected molecular chain or network. The central question is how connections form, not merely whether a material becomes solid. Joining small molecules can produce a thread that can be handled, while the individual molecules remain invisible. See IUPAC, Basic Definitions of Terms Relating to Polymers, definition 2.2.
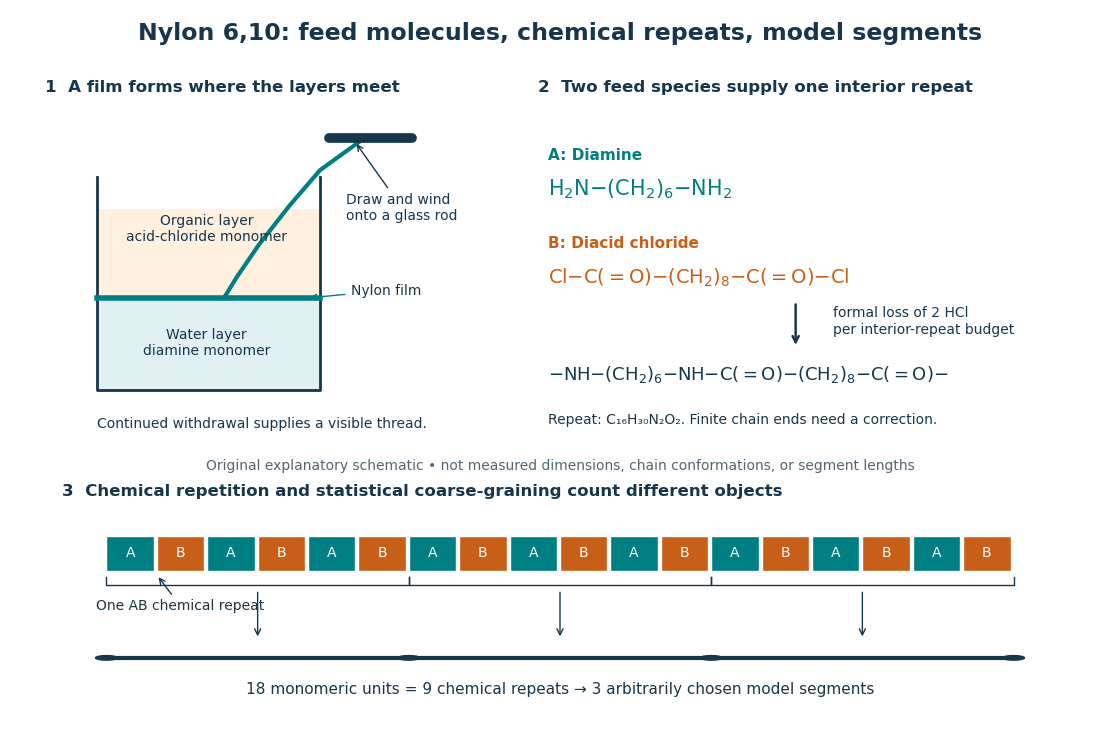
From two liquid layers to a Nylon 6,10 thread: monomers, a constitutional repeating unit, and statistical segments
. Original schematic of the authored UCSB polymerization demonstration described below. Chemical formulas identify the actual reactants and interior repeat; colored blocks identify their contributions. The lower coarse-graining sketch deliberately groups three repeats per model segment. Shapes, sizes, and that grouping are illustrative, not measured molecular geometry or a fitted Kuhn length. Details.
In step-growth polymerization, complementary reactive groups on monomers and on shorter or longer chains can react with one another. A two-unit chain can join a three-unit chain to make a five-unit chain; growth does not require a specially active end to add only free monomers. An example is the Nylon 6,10 reaction of with : each amine end can form an amide connection with an acid-chloride end, with formal elimination of . Both reactants have two suitable ends, allowing repeated joining. See Paula Hammond, MIT 10.569 lecture 2: step-growth joining and Hammond, lecture 5, p. 4: acid chloride–amine reaction.
In chain-growth polymerization, a monomer reacts with an active site on a growing chain, and the growth step regenerates an active site. Schematically, , where counts incorporated monomeric units and the star denotes reactivity rather than a particular electric charge. Ethene forming polyethylene is an example: the double chemical bond is converted into single chemical bonds as the carbon backbone grows. Initiation creates the active chains; subsequent reactions can deactivate them or transfer reactivity to another molecule. See IUPAC: chain polymerization and John McMurry, Organic Chemistry, 31.1: chain growth. Whether growth is stepwise or carried by an active site is a different question from whether small byproducts are released: chain-growth polymerization can also release them.
A motivating experiment: drawing nylon from a liquid interface. Feldwinn's UCSB demonstration places of an aqueous chemical solution of hexamethylenediamine, , in a beaker, then gently adds of sebacoyl chloride, , in an organic solvent. The procedure specifies hexane; other parts of that handout list cyclohexane. The layers remain separate. The instructor catches material at their interface with tweezers and winds it slowly onto a glass rod; the handout reports continued spooling of Nylon 6,10. It specifies freshly prepared acid-chloride chemical solution and washing the thread before touching it. These are observations and conditions of a supervised laboratory demonstration, not measurements of chain lengths. See Feldwinn, UCSB Demonstration Library, Demo 044, pp. 1–3.
The mechanistic interpretation is that the monomers encounter one another near the interface, where amide connections produce the film; withdrawing it exposes a region where more material can form. An authored research demonstration independently reports a rapidly formed interfacial film collapsing into a thread on withdrawal, using cyclohexane and an aqueous chemical solution containing potassium hydroxide. See Polymer Day: Outreach Experiments for High School Students, Journal of Chemical Education (2017), nylon demonstration. This is the striking observation: a thread much longer than the beaker can emerge because material is continually supplied to the formation region, rather than because a pre-existing thread was hidden in the liquid.
Useful proposed controls, not results reported in those demonstrations, omit one monomer at a time while retaining the same solvents, liquid amounts, and pulling procedure. The prediction is loss of a renewable drawable Nylon 6,10 film; merely finding cloudiness would not establish polymerization, since other chemical reactions, including acid-chloride hydrolysis, can occur. See McMurry, 21.4: competing acid-chloride hydrolysis. Stopping and restarting withdrawal would probe renewal at the interface. Even successful drawing does not measure the degree of polymerization, establish a chain-length probability distribution, or prove the entire growth mechanism. Chemical identification and a molecular-size measurement would be needed for those stronger conclusions. These controls and limits are inferences from the setup, not invented experimental data.
Why almost complete reaction matters in step growth. For ideal linear step-growth polymerization, assume exactly two reactive groups per starting molecule, equal numbers of complementary groups, only joining of different chains, no rings, no branching, and no loss of chain material. Let count starting monomer molecules, including both reactant species, and let be the fraction of either type of reactive group consumed. There are initially groups of each type, so connections reduce the number of chain-bearing molecules to ; the count excludes small byproducts but includes unreacted monomers. The number-average degree of polymerization, measured in monomeric units, is thereforeThus , , and give , , and . Losing the last fraction of unreacted groups matters enormously. See Hammond, lecture 3: balanced step-growth conversion. This counts residues of both feed species; a Nylon 6,10 interior constitutional repeating unit contains two such monomeric units.
For an feed with initial reactive-group counts, put and now let denote the fraction of the limiting groups consumed. Under the same linear, no-ring assumptions, and , givingFor , even complete consumption of gives . Stoichiometry controls chain length as well as yield. See Hammond, lecture 2, pp. 2–3: stoichiometric imbalance. The liquid-interface demonstration is not a homogeneous, balanced reactor, so its thread length cannot be inserted into this formula as a molecular chain length.
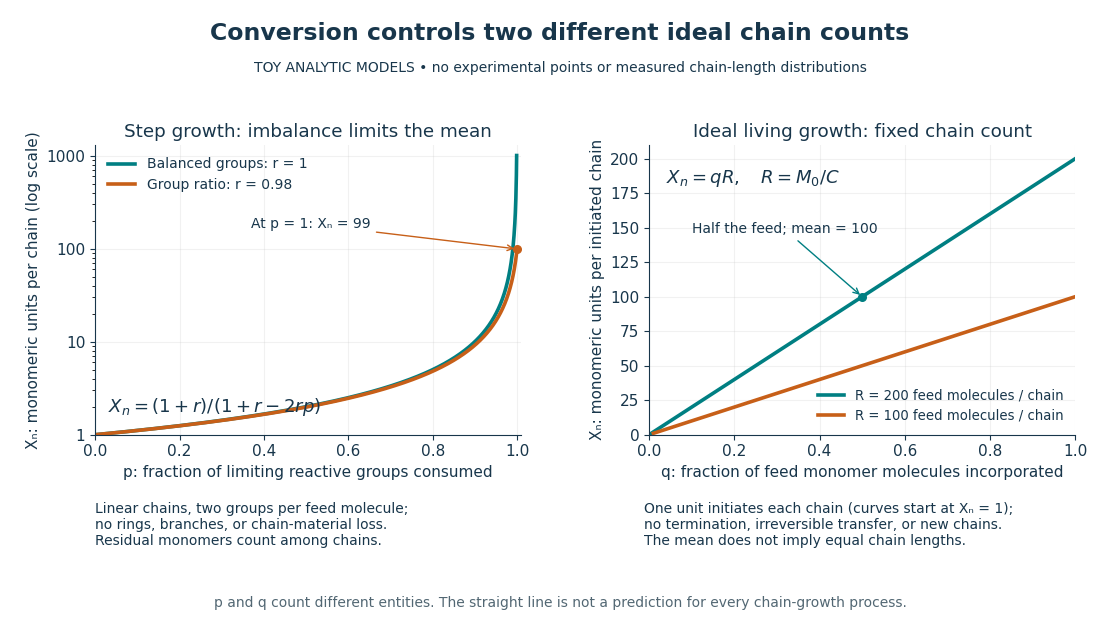
Two ideal degree of polymerization counts: late growth in step-growth polymerization and fixed-chain growth in ideal living polymerization
. Toy analytic curves, not measured data. Left: , with the limiting-group conversion and the initial reactive-group ratio. Right: , with the fraction of feed monomer molecules incorporated and their initial number per initiated chain. The two horizontal axes count different entities; the right plot represents a special ideal model, not all chain-growth polymerization. Details.
A contrasting chain count. In ideal living polymerization, assume no chain termination or irreversible chain transfer, all chains initiated before the counted growth, one chain per effective initiator, a fixed chain count , and every consumed monomer molecule incorporated into those chains. If is the initial number of feed monomer molecules and their incorporated fraction, conservation gives in monomeric units, excluding initiator fragments. For , incorporating half the feed gives . If initiation consumes one monomer molecule per chain, the fixed-chain interval starts at , rather than . See Hammond, lecture 21, p. 1: ideal living-chain count. The mean does not imply identical chain lengths. Living polymerization does not require reversible loss of monomers; the defining absence of chain termination and irreversible chain transfer is explained in Devon Shipp, IUPAC Polymer Education workshop, Radical Polymerizations II, slide 50. Ordinary chain-growth polymerization with new chains, termination, or transfer need not follow the straight line.
The useful connection is between local chemistry and a macroscopic material: selecting reactive ends and their proportions controls connectivity, while the resulting polymer's conformation is a separate problem described by models such as the freely jointed chain or Gaussian chain. Formation of chemical bonds, the count of monomeric units, and the count of statistical segments must be kept distinct.
New to topics? Read the docs here!