A chemical bond associates chemical atoms or groups of chemical atoms into a sufficiently stable structure to treat it as a distinct entity. The concept covers connections within molecules and in extended solids; see the IUPAC Gold Book, chemical bond. It is useful because composition alone does not specify how the chemical atoms are connected, and those connections help determine a chemical substance's properties and possible chemical reactions.
In a covalent bond, electrons are shared between chemical atoms. Two hydrogen chemical atoms can form a molecular hydrogen molecule, with its electrons interacting with both atomic nuclei. In the usual description of this diatomic molecule, the potential energy as a function of nuclear separation has a minimum: moving the nuclei too close or too far raises the energy. See 7.2 Covalent Bonding.
Separating bonded chemical atoms requires energy; forming the corresponding chemical bonds releases energy. Whether an overall chemical reaction releases energy depends on both the chemical bonds broken and those formed. The lines in the stoichiometry diagram indicate connections schematically, rather than literal rods or a sequence of microscopic events.
An Si–O–Si linkage. hydrolysis produces silanol groups; their condensation can regenerate an Si–O–Si linkage.
Bonding with two nodal planes containing the internuclear or coordination axis. metal and orbitals can interact with ring combinations of matching delta symmetry; symmetry alone does not imply a strong bond.
Compact valence orbitals favor strong carbon-carbon bonding, but neighboring lone-pair repulsion can weaken nitrogen-nitrogen bonds. Bond strengths depend on orbital overlap, polarity and electronic occupancy together; atom size alone is insufficient.
In a local harmonic approximation, the vibrational wavenumber obeys . Comparisons at fixed reduced mass therefore use squared frequency ratios. A local stretching force constant is not identical to a bond-dissociation energy.
Representation of an electronic structure using multiple Lewis contributors with the same nuclear connectivity. The contributors are not separate species interconverting through ordinary chemical reactions.
A hydrogen atom associated with more than one skeletal centre. Removal as a proton must be distinguished from hydride transfer; the cluster charge and electron count change differently in the two operations.
An electron-counting approach relating skeletal electron pairs to a cluster vertex count . A closo deltahedron has pairs; nido and arachno frameworks have and . The count excludes electrons in externally directed bonds or lone pairs. It is a structural guide with a defined domain, not a rule that every molecule with the same total valence count has the same shape.
The number of electrons allocated to the connected cluster framework after separating external bonds and appropriate lone pairs. For with one terminal B–H bond per vertex it is .
A pair of electrons allocated to cluster bonding. closo, nido and arachno counts distinguish the number of missing vertices relative to a closed parent polyhedron.
A metal-metal bond joins two metal centers directly through overlap of their atomic orbitals. A single bond contributes one electron to the local count at each metal in conventional electron counting. Bridging ligands can accompany direct bonding, but a short bridged separation alone does not establish its bond order.
Negative hyperconjugation is donation from an occupied lone-pair or related orbital into an antibonding sigma orbital. Nitrogen-to-P–X sigma-star donation contributes to phosphazene bonding. Short P–N bonds therefore do not require a model dominated by phosphorus pi orbitals.
Adjacent compatible p atomic orbitals allow electron density to delocalize across more than two atoms. Resonance contributors describe aspects of this shared electronic structure.
Overlap between adjacent p-type atomic orbitals permits delocalized electronic states across more than one chemical bond. A cyclic path with suitable occupancy can support aromaticity; an external lone pair can act as a donor without contributing an extra independent pair to every ring count.
An electron occupying a pi-type molecular orbital. In a conjugated planar molecule, the pi subspace is commonly built from p orbitals perpendicular to the molecular plane; its electron population excludes the sigma framework.
Alternation between shorter and longer chemical bonds along a conjugated system. In the Hückel method, unequal nearest-neighbour couplings split the frontier molecular orbitals. An electronic energy gain must be compared at a stated constraint on the mean coupling and against the geometric distortion cost.
Cohesion in a metal involving delocalized valence electrons and positive ion cores. A dispersion-only sum between neutral molecular units omits this collective electronic contribution.
A bond-dissociation energy specifies the energy needed for a stated dissociation into separated fragments. An energy from the lowest vibrational state, an energy from the bottom of a potential energy well, and a finite-temperature molar dissociation enthalpy require different conventions; do not silently mix them in an electronegativity formula.
Which fragments, in which quantum states, and at which temperature? For a neutral gaseous molecule with a specified two-electron covalent bond, the usual thermochemical convention is homolysis,Each fragment receives one of the two bonding electrons; the dots denote unpaired electrons. The final fragments are isolated from each other, with their own relaxed molecular geometry and specified electronic quantum states. The energy cost includes this relaxation, rather than merely stretching the initial chemical bond while freezing everything else. For a neutral parent, heterolysis instead produces , or the opposite assignment of electric charges; those are different final states and different energies. A value for gaseous neutral fragments cannot be transferred to ions, a chemical solution, or a solid without accounting for those changed conditions.[ref]
bond-dissociation energy. definition of bond-dissociation energy: molar enthalpy for homolysis of a specified molecular entity
The distinction is useful beyond terminology. Pauling electronegativity compares consistently defined bond-dissociation energies; photodissociation can reveal a specified ground-state threshold; a thermal chemical reaction balance requires compatible enthalpies. A threshold is also different from an activation energy: it compares initial and final quantum states, whereas a barrier can obstruct the path between them.
For a diatomic molecule, let be the electronic potential energy at bond length , with the atomic nuclei treated as fixed while finding the electronic energy. Choose zero at infinitely separated chemical atoms in the specified electronic quantum states. If the minimum occurs at , the bond potential-well depth isThe bottom of this curve describes an equilibrium bond length, not an attainable motionless vibrational quantum state. Changing the zero of potential energy shifts both endpoints equally and leaves unchanged. A quantum harmonic oscillator approximates the curve near its minimum but rises without limit, so that local approximation alone cannot supply a dissociation asymptote. A Morse potential supplies both a minimum and a finite separated-fragment limit.[ref]
dissociation energy. definition: subscripts e and 0 distinguish the potential-energy minimum and ground state
The ground-state dissociation energy starts at the lowest vibrational quantum state, with the lowest allowed rotational quantum state and the fragment states explicitly specified. For a diatomic molecule whose fragments are ground-state chemical atoms, omitting any additional rotational correction,where is the vibrational zero-point energy measured upward from the well minimum. The initial molecule already has this energy, so less additional energy is needed to reach the same final limit.[ref]
bond-dissociation energy in theoretical chemistry. definition of bond-dissociation energy in theoretical chemistry: D0 = De minus zero-point vibrational energy
For fragments that are themselves molecules, include their zero-point energies too. Define as the electronic energy difference between separately relaxed products and reactant; then, with consistent rotational conventions,Thus “subtract the parent's zero-point energy” is sufficient for atomic products but generally incomplete for molecular products. An excited final quantum state raises its channel threshold by its excitation energy; that threshold does not become the ground-product just because a detector selects it.
A bond-dissociation enthalpy is the molar enthalpy change for the specified homolysis at temperature , with reactant and products in specified standard states. Its usual units are . The superscript specifies standard states; it does not fix to room temperature.[ref]
Write for each species' molar thermal enthalpy increment. With an energy per parent molecule,Here is the Avogadro constant. The thermal increments include translational kinetic energy, molecular rotation, thermally excited molecular vibration and electronic states when populated, and the contribution to enthalpy. They are additional to the zero-point energies already included in . If values of standard enthalpy of formation at the same are available, Hess's law gives the same quantity as .
Units alone do not identify the convention. Multiplying either or by the Avogadro constant gives molar energy, but does not add thermal enthalpy. One electronvolt per dissociation corresponds to . A spectroscopic wavenumber represents energy , using the Planck constant and speed of light ; corresponds to .[ref]
Fundamental Physical Constants — Complete Listing by National Institute of Standards and Technology and others. SI defining constants h, c, e and Avogadro constant; conversions calculated from them
Take an invented diatomic molecule with a Morse potential, setting the separated-atom energy to zero:Here is an inverse length, the equilibrium bond length, and , with reduced mass . These stipulated parameters describe a simple model, not a measured chemical substance. For the one-dimensional Morse oscillator, energy eigenvalues above the minimum areConsequently,The harmonic estimate is slightly larger than this model's exact zero-point energy. The physical bound-level restriction matters: the quadratic expression is not an infinite ladder of vibrational quantum states.
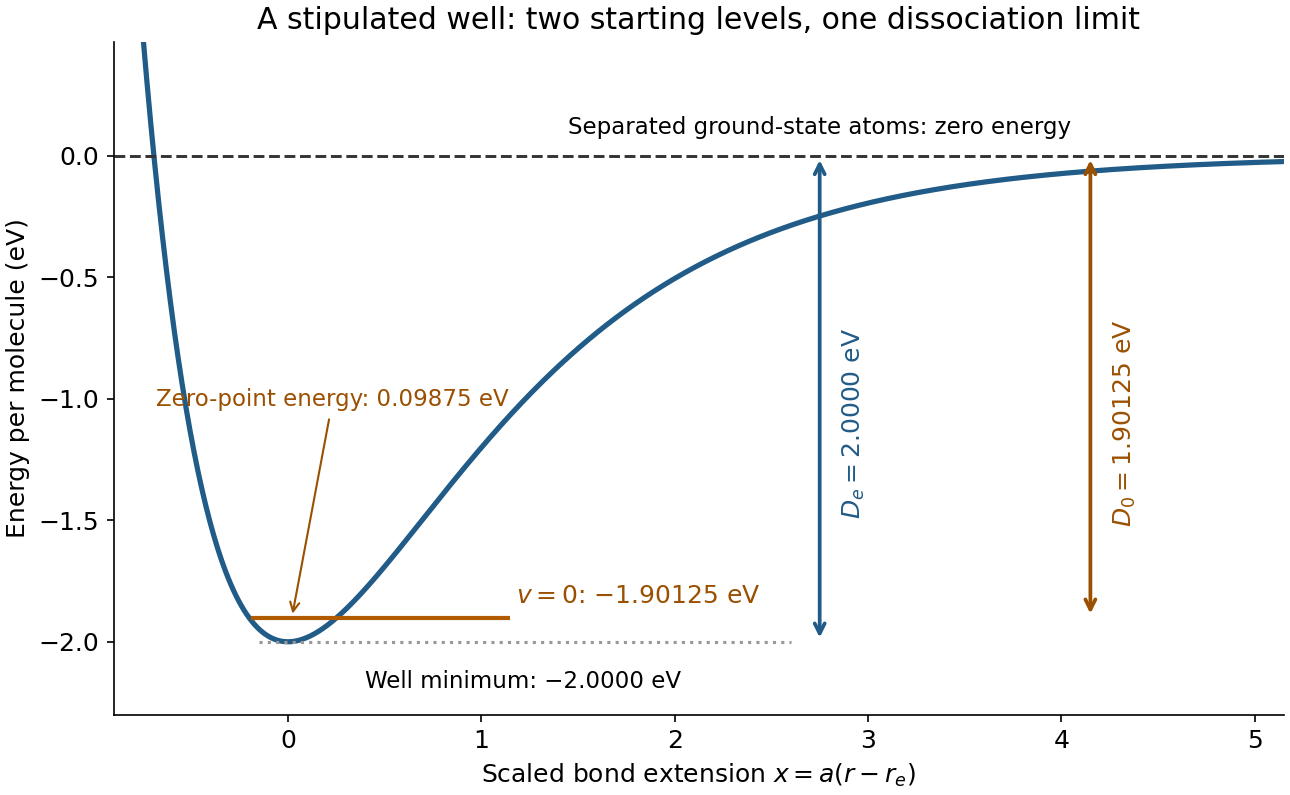
Bond potential-well depth and ground-state dissociation energy reach the same separated-atom limit from different starting energies
. Original stipulated Morse potential with and . The horizontal coordinate is ; the horizontal line is an energy eigenvalue, not a classical trajectory or a measured spectrum. Details.For a separate thermal illustration using this same stipulated , assume an ideal gas, fully active molecular rotation of the parent, negligible thermal molecular vibration, and negligible electronic excitation for all species. Each monatomic product contributes ; the diatomic molecule contributes . Thus the net increment is , with molar gas constant . At ,This last result follows from the stated thermal model, not a universal correction for every chemical bond. Frozen rotations, accessible excited states, different fragment composition, or a condensed phase change the thermal increments.
Equivalent chemical bonds in an intact molecule need not have equal successive bond-dissociation enthalpies. Once one chemical bond is removed, the next reactant has a different electronic quantum state and may relax to a different molecular geometry. The successive steps compare different species. This distinction also explains why a tabulated average chemical bond strength is not necessarily the cost of the first cleavage.[ref]
7.5 Strengths of Ionic and Covalent Bonds. “Bond Strength: Covalent Bonds”, methane paragraph: equivalent initial bonds and unequal sequential breaking costs
For an invented species , stipulate the following gas-phase steps, with all enthalpies at the same temperature and the same standard states:The symbols stand for hypothetical species with consistently chosen electronic states. They are not values for water or another measured molecule. By Hess's law, the total enthalpy of atomization for is ; the arithmetic mean per broken chemical bond is . Neither individual step equals that mean. Nor must the second step be cheaper: here it is deliberately more expensive. An exact sequence sums exactly; substituting averages taken from other molecules generally gives only an estimate.
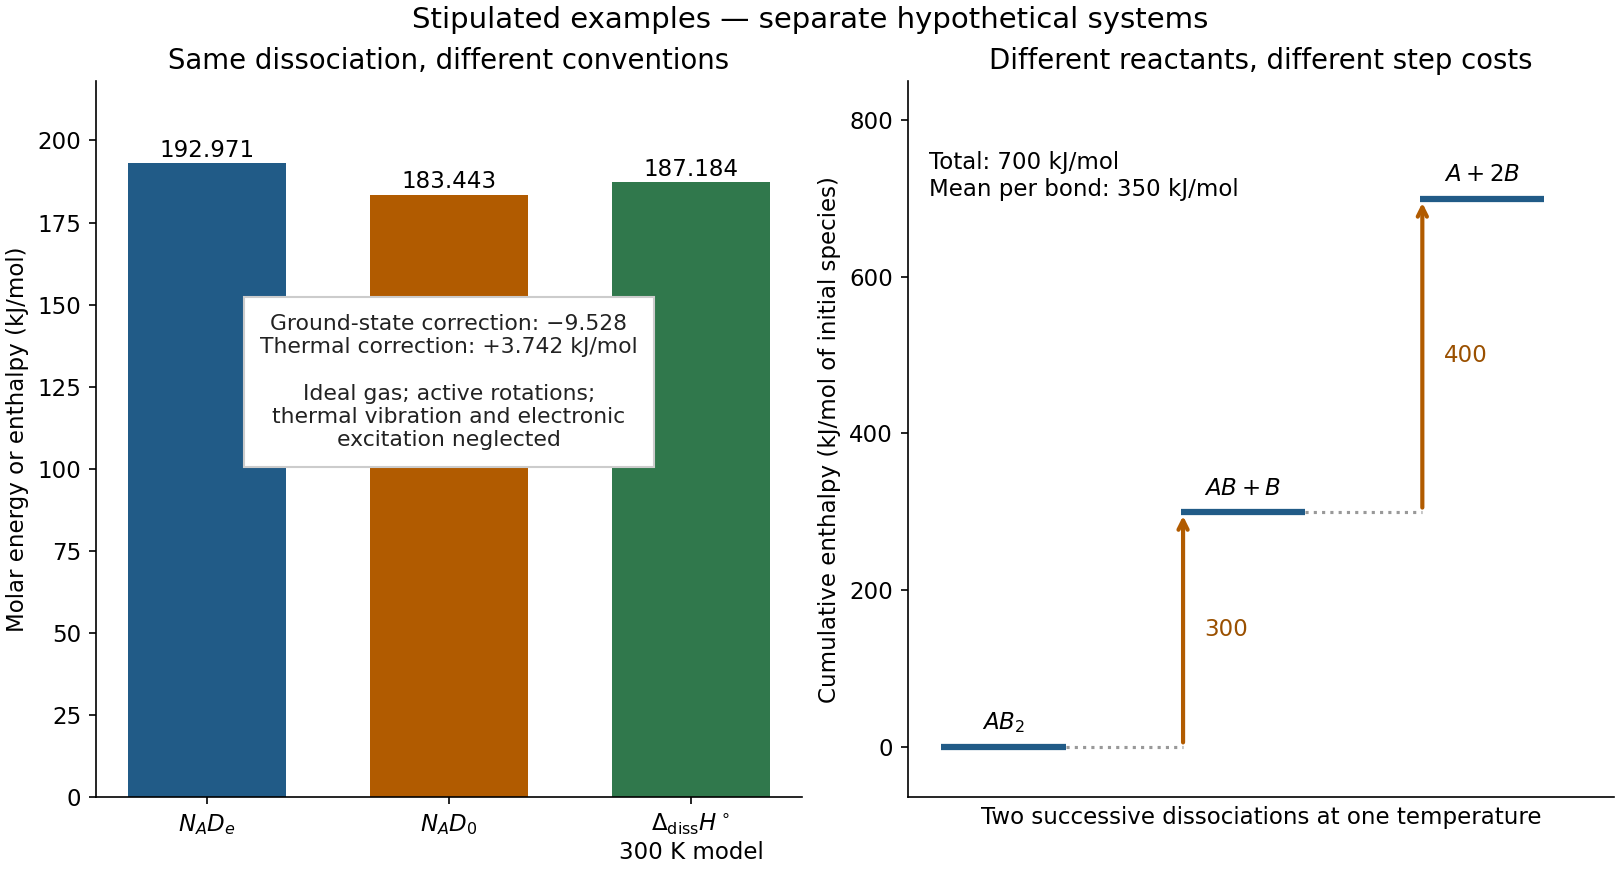
Two stipulated examples: changing the energy convention for one dissociation, and changing the reactant during sequential cleavage
. Left: , , and the model bond-dissociation enthalpy at . Right: cumulative enthalpy levels for two successive steps, with costs and and mean . The panels are separate hypothetical systems; neither contains molecular measurements. Details.Homolysis splits a covalent bond with one bonding electron assigned to each fragment. A neutral closed-shell molecule thereby produces fragments with unpaired electrons, each a radical. The usual bond-dissociation enthalpy uses this product convention.
Heterolysis assigns both bonding electrons to one fragment. For an initially neutral molecule, the resulting cation and anion define a different dissociation channel from homolysis; their isolated gas-phase energy differs from that of solvated ions.
Photodissociation separates a molecule into fragments after photon absorption. For an isolated parent with a single absorbed photon, conservation of energy givesInternal excitation energies here are relative to the parent and product ground states used to define ; is the total product translational kinetic energy in the parent's centre of mass frame. Measuring one product's kinetic energy is insufficient by itself. For two fragments of masses , conservation of momentum gives when is detected. Unknown product excitation can conceal part of the available energy.
The number of specified photochemical events per absorbed photon. A wavelength-dependent channel yield must be included when converting absorption into a chemical rate.
The first-order rate coefficient for a specified photodissociation channel. It integrates spectral actinic flux, absorption cross section and channel quantum yield. Absorption without dissociation does not contribute to that channel's rate. Its units are inverse time.
In a direct-beam model with an effective ozone cross section , removal of an overlying column multiplies the transmitted photodissociation flux by . A real photodissociation coefficient integrates wavelength-dependent cross section, quantum yield and actinic flux, including scattering and the local radiation field.
Photofragment-ion imaging detects fragment velocities by ionizing neutral products and mapping their motion onto a position-sensitive detector using an electric field. In a velocity-mapping implementation, the image radius measures transverse speed under fixed lens settings. Selecting an arrival-time slice helps extract the central plane of the three-dimensional distribution. Species identification and electronic quantum state selection are essential for turning a radius into a particular dissociation threshold.
A molecular nitrogen experiment used a pulsed nozzle and two skimmers. A tube raised the rotational temperature. Opposed vacuum-ultraviolet laser pulses excited selected molecular levels and, about later, selectively ionized excited nitrogen fragments. A microchannel plate detector, phosphor screen and camera recorded radii.[ref]
Accurate measurements of the bond dissociation energies of 14N2, 14N15N and 15N2 by Liya Lu and others. Lu, Jiang and Gao, §2, printed manuscript pp. 2–3: nozzle, tube, pump/probe and imaging readout
The measured squared radii were fitted against molecular excitation energies, referenced to the parent ground state:The intercept at zero radius gave the excited-product threshold; the fitted slope absorbed the detector scale. A wavemeter monitored the excitation source; scanning the probe covered the recoil Doppler shift. Subtracting the atomic excitation energies converted the intercept to ground-product .[ref]
Accurate measurements of the bond dissociation energies of 14N2, 14N15N and 15N2 by Liya Lu and others. §1: image-radius equation; §2: wavemeter and probe scan; §3 and Figure 4: fits; §4: atomic excitation correction
An apertured grounded plate and detector timing rejected faster competing channels. Selecting fragment electronic states distinguished nearby thresholds. Agreement of across isotopes after zero-point energy correction tested consistency. The method still requires correctly assigned levels, adequate near-threshold signal and stable imaging; a signal drop alone can reflect another channel or a barrier.[ref]
Accurate measurements of the bond dissociation energies of 14N2, 14N15N and 15N2 by Liya Lu and others. §2: shielding plate and timing; §3: state-selected channels; §4 and Table 2: isotope consistency and limitations
This measurement determines a state-resolved ground-state dissociation energy. Obtaining a finite-temperature bond-dissociation enthalpy requires the thermal increments above; obtaining a bond potential-well depth requires the zero-point energy correction. The three quantities describe different starting conditions for the same specified separation.
Electronegativity describes a chemical atom's tendency to attract electron density towards itself in a chemical bond. It helps answer a useful question: when two different chemical atoms share electrons, which end of the chemical bond tends to be richer in electron density? For hydrogen chloride, the answer is chlorine: the covalent bond is a polar covalent bond, conventionally written . The symbols indicate partial atomic charges, not isolated ions carrying whole elementary electric charges.[ref]
Electronegativity is a comparison scheme, not the number of electrons on a chemical atom. A partial atomic charge depends on how the electron density is partitioned; formal charge instead follows an integer allocation in a Lewis structure. Neither is numerically equal to electronegativity. Nor is electronegativity identical to electron affinity, which concerns adding an electron to an isolated species. Different electronegativity scales use different measured inputs and conventions; compare numbers only after specifying the scale.[ref]
For a concrete comparison, the Royal Society of Chemistry lists Pauling electronegativities and . Their difference is , so the electronegativity rule predicts bond polarization towards chlorine in hydrogen chloride. An isolated molecular hydrogen molecule has identical ends and zero electronegativity difference. These examples explain a direction of unequal sharing, not a quantitative partial atomic charge.[ref][ref]
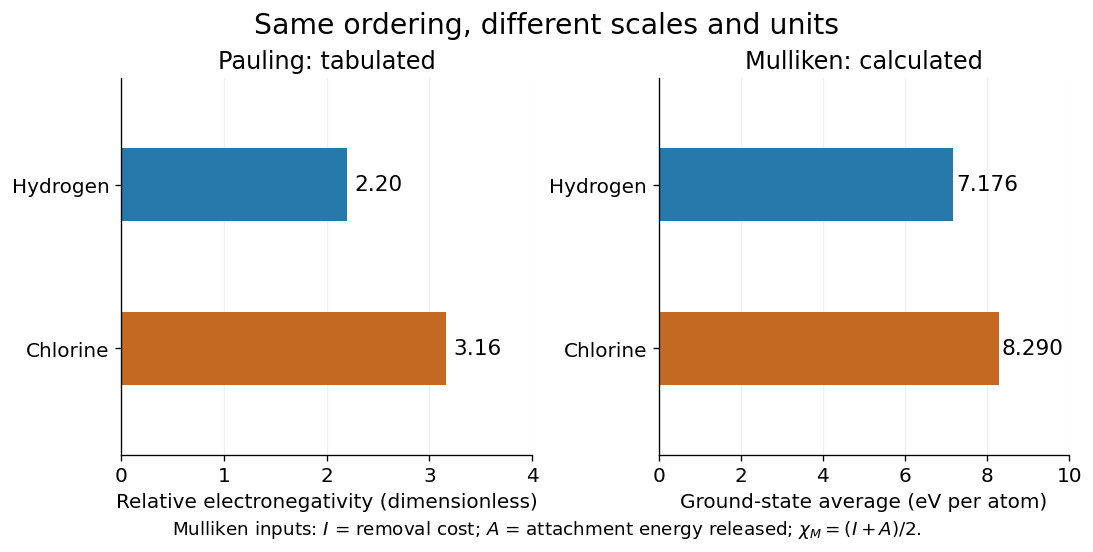
Pauling electronegativity and Mulliken electronegativity for hydrogen and chlorine: separate scales, separate units; the Mulliken electronegativity values are calculated from the stated ground-state inputs
. Details. A larger electronegativity difference often suggests stronger bond polarization, but there is no universal numerical boundary where a covalent bond suddenly becomes ionic bonding. Tables of cutoffs are classification aids with exceptions. Hydrogen fluoride, for example, is described by a polar covalent bond even when its electronegativity difference exceeds a textbook's nominal ionic threshold. The actual electron density, physical setting, and chemical substance's structure matter.[ref]
Chemistry 2e. §7.2, “Electronegativity and Bond Type”, general-guide warning and HF exception
The distinction from ionization energy is equally useful: an ionization energy measures the cost of removing an electron from a specified isolated species, whereas electronegativity compares attraction for electron density in a bonded environment. Neither the removal cost alone nor the attachment release alone supplies a unique bonded-atom electronegativity. A scale combines selected measured inputs with a model and a convention.[ref]
electronegativity. E01990: multiple definitions; Mulliken average versus Pauling bond-energy differences
For an original relative comparison, stipulate fictitious bonded sites on one chosen dimensionless electronegativity scale:The electronegativity heuristic predicts bond polarization towards in , towards in , and towards in . Shifting leaves every comparison unchanged. Although the difference is twice the difference, this does not make its partial atomic charge or bond dipole twice as large: those require additional information about electron density and bond length. These symbols describe a stipulated ordering, not a table of measured chemical elements.
Keep the chemical context attached to that ordering. A chemical atom in a molecule has neighbours and an electron configuration; an isolated-atom energy scale and a bonded-atom comparison are different constructions. The chemical environment dependence of electronegativity explains why a table is a useful approximation rather than an immutable property of every site. Furthermore, an applied electric field can induce bond polarization even between identical chemical atoms, through electric polarizability; equal tabulated electronegativity does not forbid that response.
A scale proportional to the average spectroscopic one-electron binding energy of a free ground-state atom’s valence electrons. For an s,p atom, , with positive binding energies and a scale constant . It distinguishes s and p binding rather than assigning both one Slater charge.[ref]
Pauling electronegativity is a relative, dimensionless scale motivated by bond-dissociation energies. The interesting observation is that a chemical bond between unlike chemical atoms can be stronger than a simple baseline inferred from the corresponding like-atom chemical bonds. An empirical excess energy can then be associated with an electronegativity difference. The 1932 paper The Nature of the Chemical Bond. IV. The Energy of Single Bonds and the Relative Electronegativity of Atoms built such a map; its original coordinates used hydrogen as zero, so those coordinates should not be mistaken for the modern tabulated Pauling electronegativities.[ref]
One arithmetic-baseline version of the Pauling scale relation isHere , and are consistently defined bond-dissociation energies per molecule, and the arithmetic mean is the chosen baseline. With molar energies instead, write , approximately . The denominator makes the result dimensionless; the factor is not a unit-independent magic constant. This is an empirical relation with a specified baseline, not a definition valid for every chemical bond.[ref]
Energy Electronegativity and Chemical Bonding by Stepan S. Batsanov. §1 equation (1), coefficients for eV, kcal/mol and kJ/mol; §2.2, dependence on other bond properties
For an explicitly synthetic example, suppose and with the same temperature and dissociation convention. Then and the model gives an electronegativity difference of . Replacing by gives zero; a negative excess would make this real square-root model inapplicable, not produce a negative electronegativity difference. These invented inputs illustrate the formula and are not measurements of named chemical substances.
Only differences appear, so adding the same constant to every Pauling electronegativity leaves this relation unchanged. A reference value must fix the origin. The bond-dissociation energies alone also do not tell which endpoint attracts electron density more strongly, because interchanging and leaves the formula unchanged. Chemical comparisons fix that direction. Thus Pauling electronegativity is useful as a compact ordering, while the underlying measured energies and approximations remain essential.
A Pauling bond-energy excess subtracts a chosen like-atom baseline from the unlike-atom bond-dissociation energy. For an unequal-input arithmetic example, stipulate , and , all with one dissociation convention. The arithmetic mean baseline and excess areThe arithmetic-baseline Pauling scale model therefore gives . The bond-dissociation energy is not itself the excess: subtracting the baseline is essential. All three numbers are stipulated, not experimental values for named chemical elements.
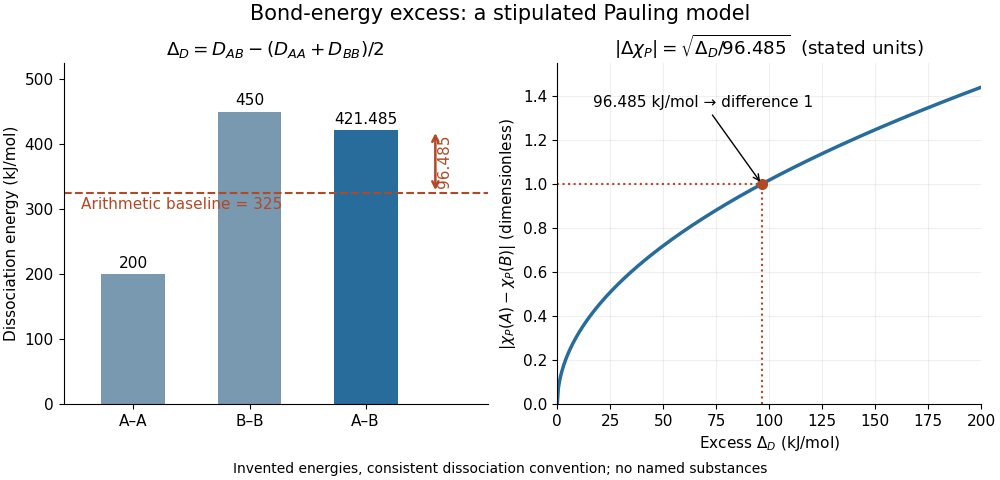
Constructing a Pauling bond-energy excess from stipulated bond-dissociation energies and converting it to a dimensionless electronegativity difference
. Details. Changing the baseline changes the model. For the same stipulated inputs, a geometric mean baseline would be , yielding an excess of and a difference of approximately . It is a different construction, even though both start from the same bond-dissociation energies. State which baseline is used before comparing electronegativity differences.
The arithmetic-baseline Pauling scale also has a useful inverse: a difference of corresponds to an excess of , and a difference of corresponds to . Doubling the electronegativity difference quadruples the modeled excess. Interchanging the endpoint labels changes neither quantity; the square root alone cannot choose the direction of bond polarization.
In The Nature of the Chemical Bond. IV. The Energy of Single Bonds and the Relative Electronegativity of Atoms (printed pp. 3576–3578, Table II and equation (1)), Linus Pauling organized excess energies for chemical bonds into an approximately additive map of their square roots. He also identified a limitation: his H–F map separation of predicted an excess of , exceeding his table's value by . Those are historical values and a historical model discrepancy, not modern recommended inputs.[ref]
The Nature of the Chemical Bond. IV. The Energy of Single Bonds and the Relative Electronegativity of Atoms by Linus Pauling. printed pp. 3576–3578, Table II, equation (1), and paragraph beginning “These coördinates”
Mulliken electronegativity uses both the cost of removing an electron and the energy released by accepting an electron. For specified isolated-atom states, the common ground-state convention isThe free electron's energy is the zero reference. Here is the ionization energy and is the electron affinity, positive for exothermic attachment. If attachment is instead tabulated as the signed energy change , the same expression is . Stating the sign convention prevents adding the wrong quantities. The 1934 original treated suitable valence states, which are not automatically the ground states used in this example.[ref]
Mulliken electronegativity has energy units before any additional rescaling. With and in electronvolts per chemical atom, is in electronvolts; with both in , it is a molar energy. For hydrogen, the RSC ground-state inputs are and , giving per chemical atom. For chlorine, and give . These are calculated ground-state Mulliken electronegativities, not the RSC's tabulated Pauling electronegativities.[ref][ref]
Both schemes place chlorine above hydrogen here, but their numerical differences cannot be subtracted from one another or equated. Dividing a molar Mulliken electronegativity by an energy conversion factor changes units; it does not transform it into a Pauling electronegativity. A further empirical mapping would require its own calibration and stated domain.
Electron affinity measures how strongly an isolated neutral species binds an additional electron. In the spectroscopic convention used here, it is the positive energy required to detach that electron from a bound anion. With the free electron at rest assigned zero energy,The quantum states of and must be specified, usually their ground states. Positive means that attachment lowers the energy; a signed attachment energy is then negative. This convention agrees with the IUPAC definition and with the atomic tables used for Mulliken electronegativity.[ref]
For chlorine, a published photodetachment threshold is , approximately per chemical atom. Here is the Planck constant, the speed of light, and the inverse-length unit expresses an energy through a spectroscopic wavenumber. The parentheses give the reported uncertainty in the last digits, not a range of possible sign conventions.[ref]
Isotope shift in the electron affinity of chlorine by U. Berzinsh and others. §2.3, printed pp. 6–7: reported threshold 29138.59(22) inverse centimetres
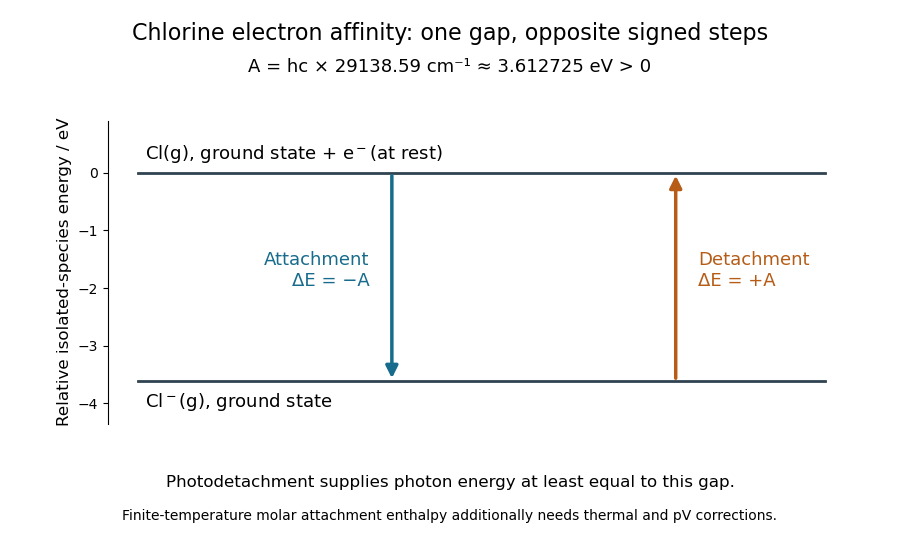
Electron affinity of chlorine: positive detachment energy and negative attachment energy connect the same isolated-species levels; a finite-temperature electron attachment enthalpy needs additional corrections
. Details. An electron affinity is neither an electronegativity nor a lattice energy. The first describes a specified isolated-species energy difference, electronegativity compares attraction for electron density in a chemical bond, and lattice energy refers to an entire ionic crystal and separated gaseous ions. A stable anion can therefore help explain ionic bonding without describing the full energy balance for making a salt.
An electron attachment enthalpy is the signed molar enthalpy change for a stated attachment reaction,For favourable attachment such as chlorine's first attachment, the electron attachment enthalpy is negative. Its leading isolated-ground-state energy contribution is , where is the Avogadro constant. At finite temperature, consistent thermal populations, translational contributions and the term in enthalpy must also be included, together with the same gaseous-electron reference used in the corresponding ionization energy term. Consequently a measured spectroscopic electron affinity is not automatically an exact attachment enthalpy with a minus sign. A Born-Haber cycle must use compatible enthalpies throughout.
Photodetachment removes an electron from an anion using a photon. For chlorine, the reaction is . Measuring the minimum photon energy for the specified final quantum state gives a photodetachment threshold and hence an electron affinity.
In the published chlorine experiment, negative ions were accelerated and mass-selected, then overlapped with a tunable laser beam. A downstream electric field deflected charged ions; neutral chemical atoms continued to a plate, whose secondary electrons were counted. Scanning the laser frequency produced an onset in this neutral-atom signal. The apparatus therefore inferred an electron affinity from a threshold, rather than measuring the heat of attachment.[ref]
Isotope shift in the electron affinity of chlorine by U. Berzinsh and others. §2, printed pp. 2–4, Figure 1: ion preparation, deflectors, neutral-atom secondary-emission detector and threshold fitting
Reversing the laser direction and averaging the two thresholds corrected the first-order Doppler shift. A Fabry–Pérot interferometer and reference spectral lines calibrated the frequency. Finite laser bandwidth, unresolved quantum state splitting and shifts of the calibration lines limited the inference; the authors included these effects in fitting and the error estimate. The level diagram shows only the extracted gap, not a measured signal curve.[ref]
The contrast between Pauling electronegativity and Mulliken electronegativity becomes clearer by following the measurements into the formulas. A Pauling bond-energy excess compares three compatible bond-dissociation energies; Mulliken electronegativity takes the arithmetic mean of an ionization energy and an electron affinity. The former asks how much stronger an unlike-atom chemical bond is than a stipulated baseline; the latter combines two different isolated-species energy gaps.
Actual readouts precede those model choices. In the nitrogen dissociation threshold from fragment images, fitted squared image radii versus assigned excitation energies reveal a threshold after accounting for the product quantum states. In the chlorine photodetachment threshold experiment, mass-selected anions overlap a tunable laser, and neutral-atom counts reveal the onset of electron detachment. Opposite laser directions control the Doppler shift. That second threshold supplies an electron affinity, not an electronegativity; the Mulliken scale still needs an independent ionization energy.[ref]
Accurate measurements of the bond dissociation energies of 14N2, 14N15N and 15N2 by Liya Lu and others. §1, recoil-energy/image-radius equation; §3.2, Figure 5, squared image sizes versus term energies; §3.2, threshold channel assignment
[ref]For an original arithmetic comparison, stipulate isolated-species and positive attachment-release for fictitious species . Their Mulliken electronegativities are . The largest ionization energy belongs to , while the largest Mulliken electronegativity belongs to : accepting an electron matters as well as retaining one. Using signed attachment energies in gives exactly the same results.
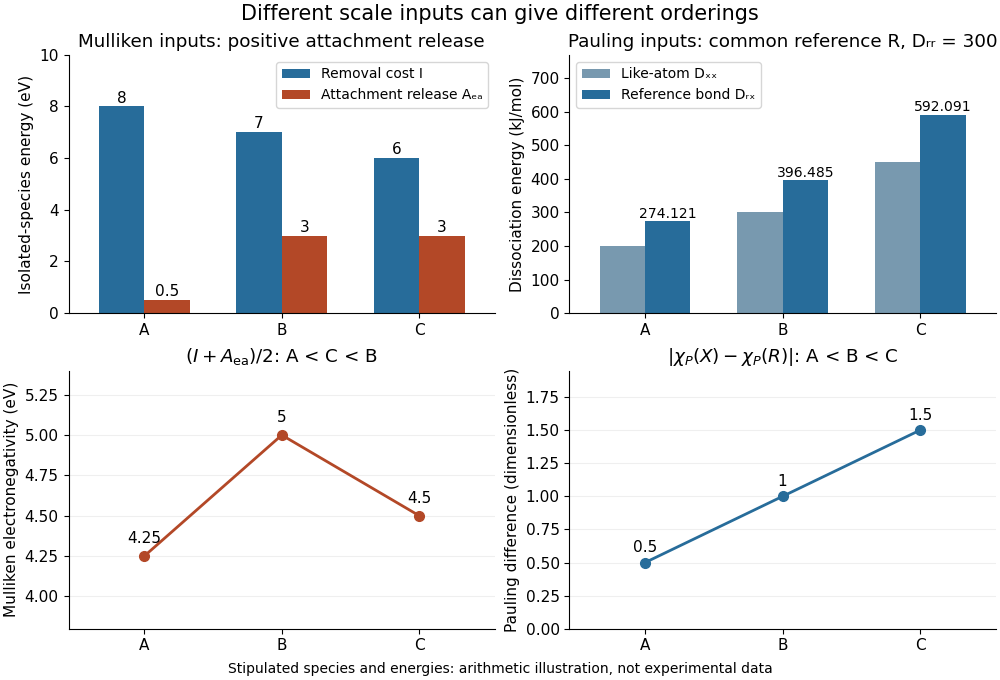
Stipulated inputs to Mulliken electronegativity and arithmetic-baseline Pauling electronegativity: different energies, units and orderings
. Details. Independently stipulate and for , with unlike-atom valuesThus , and the Pauling scale differences from are . If all three fictitious species are placed above on that scale, their ordering is , whereas the stipulated Mulliken scale gives . The bond-dissociation energies and isolated-species energies were invented independently: this demonstrates what the formulas do, not a measured disagreement for actual chemical elements. A unit conversion cannot force these two input models to agree.
A tabulated electronegativity is useful because it compresses many comparisons, but one number per chemical element suppresses chemical context. Changing the bonded partners, bond length or electron configuration changes the conditions under which a chemical atom attracts electron density. The bond-dissociation energy can change for reasons beyond bond polarization, so attributing every extra unit of energy solely to an electronegativity difference is an approximation.[ref]
Energy Electronegativity and Chemical Bonding by Stepan S. Batsanov. §2.2, opening paragraph: bond distance and atomic valence affect bond energy
An oxidation state labels a formal allocation of electrons; it is not a measured partial atomic charge. State-sensitive electronegativity models may give different values for different oxidation states or electron configurations, while a simple chemical element table intentionally provides a common comparison convention. Likewise, replacing an isolated chemical atom by a site in an ionic crystal introduces a surrounding electric field and many neighbours. Applying the same tabulated electronegativity without adjustment becomes an additional assumption.[ref]
For the ground-state Mulliken scale, the state dependence is visible directly in the formula. Replacing by replaces with as the removal cost, and replaces with as the attachment release. These are different ionization energies and electron affinities. This isolated-ion arithmetic does not claim that a bonded chemical atom with the same formal oxidation state is an isolated ion. In a molecule, the actual electron density and dipole evidence for bond polarization supply complementary information about that environment.
A permanent electric dipole moment gives evidence of separated positive and negative electric charge in an overall neutral molecule. This is a more direct probe of chemical polarity than an electronegativity table. For hydrogen chloride, NIST lists about debye for the lowest vibrational state and debye for the equilibrium value. They describe different averaging conventions, not two contradictory measurements. The NIST WebBook traces the vibrational value debye to molecular-beam electric-resonance measurements in 1970.[ref][ref]
How does molecular beam electric resonance reveal an electric dipole moment? A source and apertures form a beam of molecules in vacuum. A first nonuniform electric field, A, selects quantum states through their different Stark effects. In a central region, C, electrodes supply a nearly uniform electric field and applied radiation drives transitions at selected frequencies. A second nonuniform electric field, B, analyzes the outgoing quantum states; a detector measures the rate of arriving molecules. In a transmission-loss arrangement, resonant radiation transfers some molecules to a quantum state rejected by B, so the measured arrival rate dips. The observable is a change in detected molecules as the radiation frequency is scanned, rather than light absorption itself. This general apparatus description is documented in a later experimental thesis; it is not a reconstruction of the exact 1970 HCl instrument.[ref]
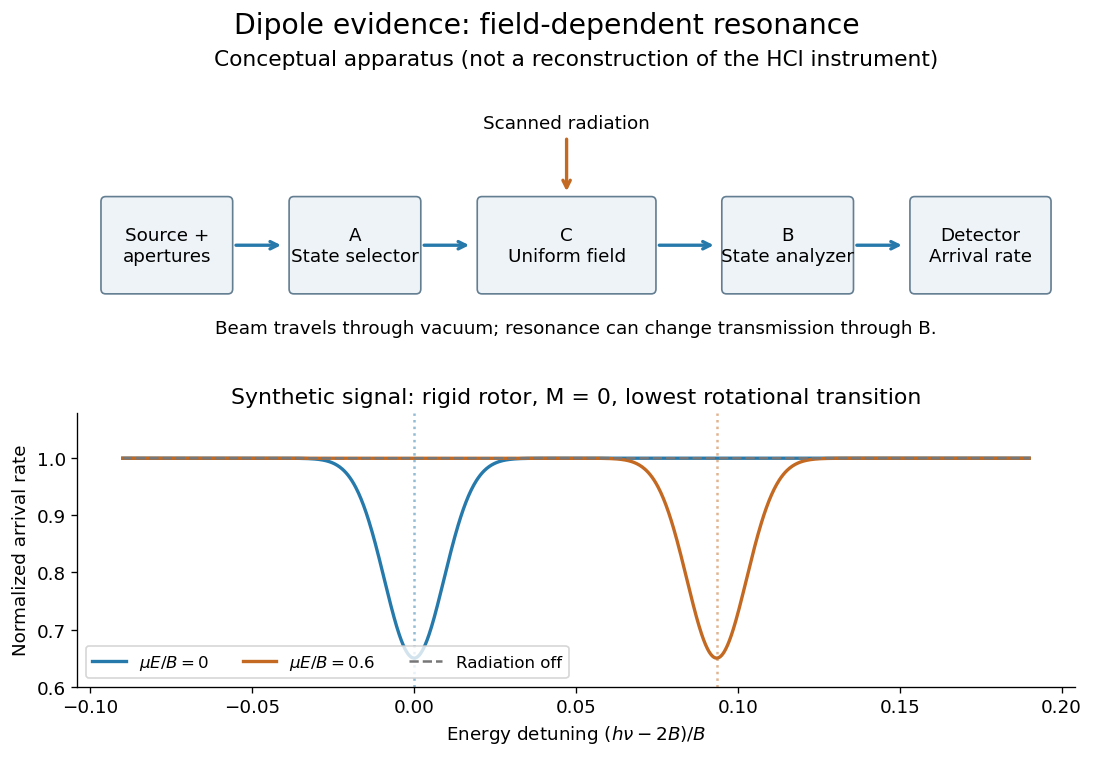
Molecular beam electric resonance: schematic state selection and a synthetic Stark effect signal from a rigid rotor model; neither apparatus dimensions nor signal curves are measured HCl data
. Details. The inference comes from how transition frequencies change with the known electric field. The interaction couples the electric dipole moment to the electric field; fitting the quantum mechanics of the relevant rotational states extracts its magnitude. A free rotating molecule need not show a first-order shift proportional to : the weak-field Stark effect can be quadratic. The diagram uses a simplified rigid rotor, with an energy constant, to illustrate this behavior. It omits the fine details of actual spectra.
Useful controls include scans with the radiation off to measure the baseline arrival rate, repeated scans at several calibrated electric fields, and agreement between independently assigned transitions. These are methodological checks, not claims about undocumented steps of the 1970 experiment. Field calibration and state assignment limit the inferred electric dipole moment. The published 1970 abstract confirms analysis of HCl and DCl beam-resonance spectra and combination with infrared spectroscopy intensities to obtain a distance-dependent electric dipole moment; its full apparatus and calibration record were unavailable here.[ref]
Dipole Moment and Hyperfine Parameters of H35Cl and D35Cl by E. W. Kaiser. pp. 1686–1703, authored abstract; full text unavailable
Even an accurately measured electric dipole moment is not an absolute partial atomic charge. Replacing a hydrogen chloride molecule by two point electric charges separated by bond length gives . Using debye and gives times the elementary electric charge. This is an effective two-point model, combining a vibrationally averaged electric dipole moment with an equilibrium bond length, not a unique partition of the actual electron density or a measured “18.1 percent ionic bond”.[ref]
For a molecule with several chemical bonds, molecular geometry can cancel opposing contributions to the total electric dipole moment. Carbon dioxide illustrates why an overall nonpolar molecule need not have unpolarized individual chemical bonds. An electronegativity comparison helps anticipate local bond polarization; the measured molecular electric dipole moment tests the entire electric charge distribution.
The apparatus and dipole inference are described under dipole evidence for bond polarization. The rigid rotor supplies a general starting model for rotational quantum states; rotational spectroscopy measures their transition frequencies, while molecular beam electric resonance can select and analyze states through their Stark effects.
An actual implementation gives substance to the apparatus description above. In Johannes Jacobus van Vaals's experiments, heated samples expanded with argon through a nozzle, then molecular beam skimmers formed the molecular beam in vacuum. Nonuniform electric fields selected and analyzed quantum states; parallel plates housed the microwave interaction. A surface ionization detector supplied electric current to an electronic amplifier and a lock-in amplifier. Resonant transfer into a rejected quantum state reduced the detected flux. Phase locking stabilized the microwave frequencies; an absorber suppressed reflections that could distort spectral lines. The thesis reports preliminary electric dipole moment components for gaseous potassium cyanide from measurements in nonzero electric fields. Such inference uses the Stark effect, rather than measuring electronegativity. The source notes restrictions from electric-dipole selection rules and molecular beam transit time; its preliminary dipole report is not a full calibration record. These are documented features of this apparatus, not proposed controls for the earlier hydrogen chloride experiment.[ref]
Bond polarization is unequal sharing of electron density across a chemical bond. An electronegativity difference is a heuristic for its direction and tendency, not a unique partial atomic charge.
A simple two-site model makes the notation explicit. Represent each site by effective positive electric charge , and allocate expected electron populations to and to , with . Here is the elementary charge; these effective sites omit the internal structure of real chemical atoms. Subtraction givesFor the stipulated illustration , the populations are and , and the effective partial atomic charges are and . Fractional expected populations do not split an individual electron into pieces. The total population remains two, and the pair remains neutral. No formula connecting to has been assumed; many electron density distributions and partitioning conventions can share one qualitative electronegativity ordering.
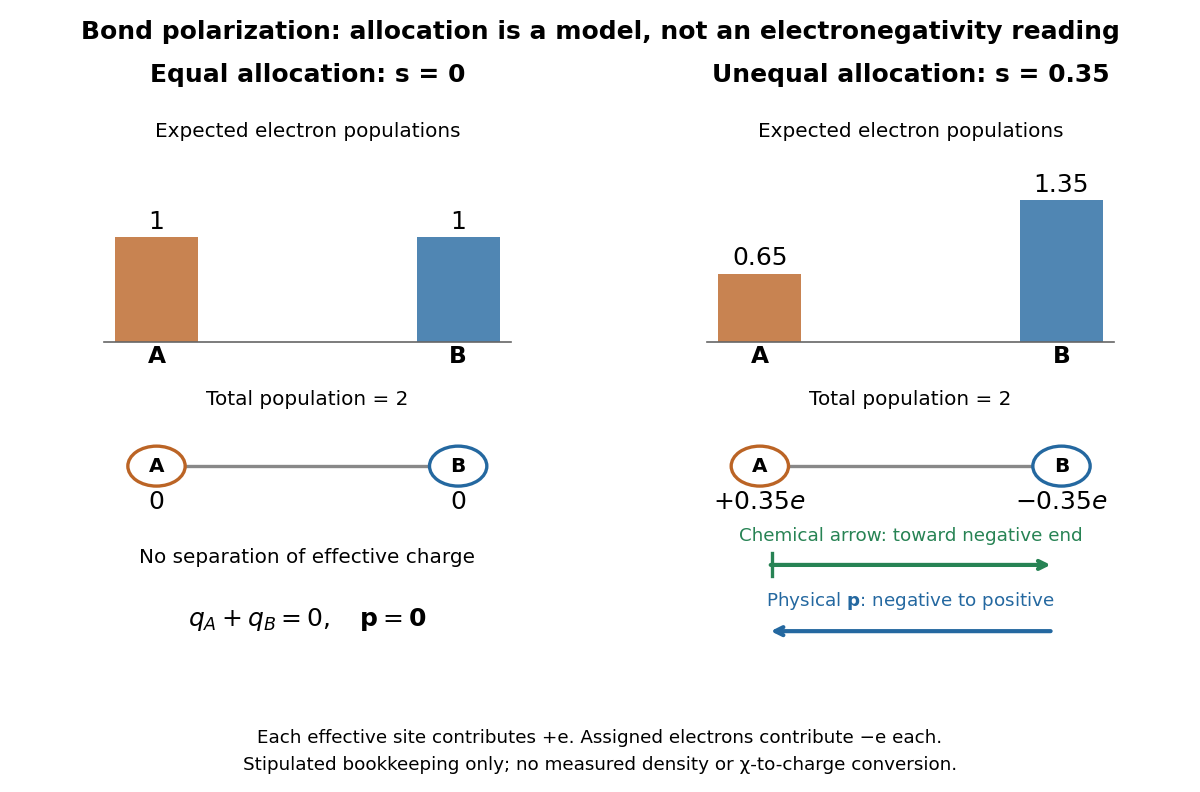
Stipulated two-site bond polarization: expected electron populations, effective partial atomic charges, and opposite chemical and physical arrow conventions
. Original model with and , effective positive site charges , and two expected electrons in total. The population bars are bookkeeping weights, not a measured electron density map. Arrow lengths illustrate direction only; no empirical electronegativity values or charge calibration are implied. Details.A bond dipole represents the separation of positive and negative electric charge associated with a chemical bond. In a neutral two-point charge model put at and at , with . Its physical electric dipole moment isIf is the negative end, the physical electric dipole moment points from to . The customary crossed-tail chemical arrow points towards the negative, more electronegative end, from to . It therefore has the opposite direction. State the convention before doing vector addition; use physical electric dipole moments in the interaction potential energy , where is the electric field.[ref]
For a fixed neutral molecule, its complete electric dipole moment follows from all atomic nuclei and the electron density, not just point charges assigned to sites. If is the expected number of electrons per unit volume, the corresponding charge density is ; the nuclear terms contribute separately. Partitioning that distribution into bond dipoles is a model. Lone pairs and electric polarization within a chemical atom can contribute as well. A measured molecular electric dipole moment cannot uniquely recover every individual bond dipole or partial atomic charge.
Molecular dipole cancellation occurs when nonzero contributions to a molecule's electric dipole moment cancel under vector addition. In a bond dipole model, use ; adding magnitudes instead discards the molecular geometry. A symmetric linear molecule with two equivalent opposed bond dipoles has and , hence zero total electric dipole moment even though each chemical bond is polarized. Carbon dioxide provides the familiar molecular example; water's bent molecular geometry permits a nonzero total.[ref]
Chemistry 2e. §7.6, Figure 7.27 and accompanying paragraphs: cancellation in carbon dioxide, noncancellation in water
For an original geometry calculation, give a fictitious three-site model a central effective electric charge at the origin and two terminal electric charges atHere is the stipulated bond length, the angle between the two connections, and is independent of any electronegativity scale. Equivalently, split the central into two colocated contributions, one for each neutral bond dipole. With , the physical electric dipole moments point from the terminal sites towards the central site:The sideways components cancel; the components along the angle bisector add. Thus . At the result is zero; at the stipulated it is exactly . This is an angle example for a model, not the measured angle or electric dipole moment of water.
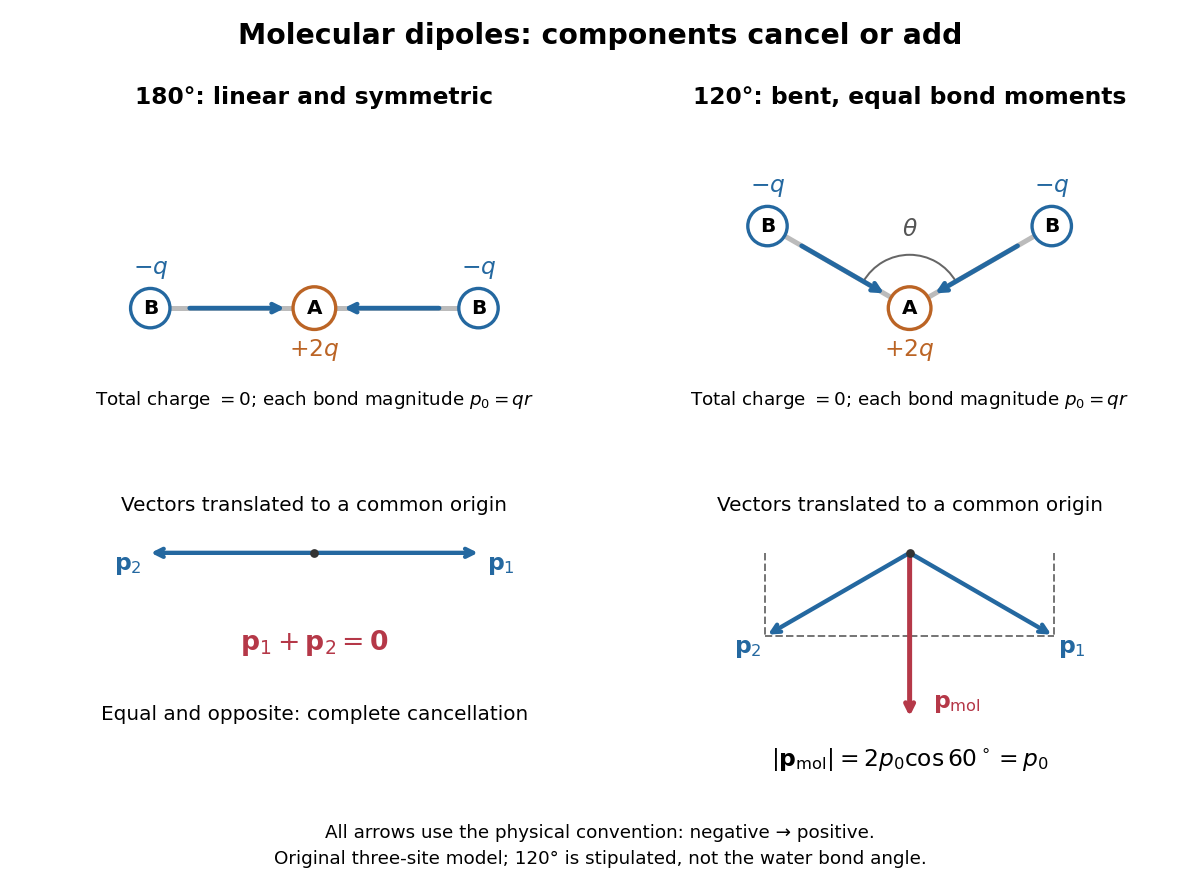
Vector addition of physical bond dipoles in a stipulated neutral three-site model: cancellation at , a bisector resultant at
. Central effective electric charge , terminal electric charges , and equal bond lengths . Arrows point from negative to positive electric charge. The lower panels translate the bond dipoles to a common origin and show their components; they are vectors, not electron trajectories or extra bonds. Details.Opposition alone is insufficient: a linear molecule with unequal opposed bond dipoles of magnitudes and retains magnitude . Conversely, symmetry can enforce exact cancellation beyond a particular bond dipole partition. If the complete charge density invariant under point reflection about the origin satisfies , contributions to cancel in pairs. That proof uses the whole electric charge distribution, including lone pairs and atomic nuclei. It explains why molecular symmetry is stronger evidence for zero permanent electric dipole moment than a rule based only on tabulated electronegativity differences.
A permanent molecular electric dipole moment is specified relative to the molecule's orientation. An ensemble with random orientations can have zero mean laboratory vector while each molecule is polar. That orientational average differs from internal molecular dipole cancellation. The Stark effect and molecular beam electric resonance probe molecular electric dipole moments through state-dependent responses to an electric field.
The word “polar” qualifies the sharing within a covalent bond; it does not require complete transfer of an electron or an isolated cation–anion pair. In , the signs identify the relatively electron-poor and electron-rich ends, while leaves the amount unspecified. A neutral Lewis structure can give both chemical atoms zero formal charge and still correspond to a polar covalent bond: equal division of the bond pair is a formal allocation, whereas bond polarization concerns the actual electron density.
Local and molecular descriptions answer different questions. Calling one chemical bond a polar covalent bond does not establish that the whole molecule is a polar molecule. Combine the bond dipoles with the molecular geometry, then check for molecular dipole cancellation. This lets a short electronegativity comparison explain a local tendency without confusing it with the observable electric dipole moment of the whole molecule.
Ionic bonding binds oppositely charged ions through attraction governed by electrostatics. In ordinary solid sodium chloride, each cation interacts with many anions, and also with other cations: attraction and repulsion together contribute to the binding of an extended ionic crystal. The chemical formula specifies equal numbers of the two kinds of ions; it does not divide the solid into isolated two-atom molecules.[ref]
This distinction explains a striking combination of properties: ordinary dry sodium chloride conducts electric current poorly, while its molten form or a sufficiently concentrated chemical solution in water conducts well. The ions already exist in the solid; their ability to move through the sample changes. A conductivity experiment on salt solutions tests this transport picture, rather than simply asserting that charged constituents must make every material conductive.
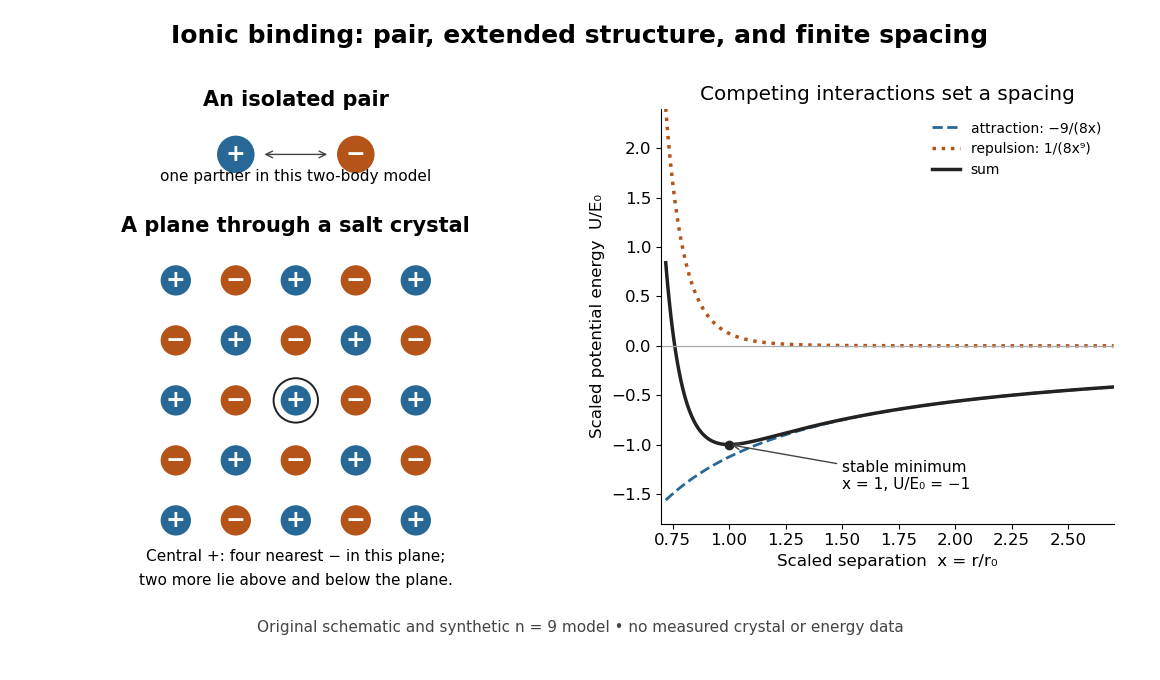
An isolated ion pair, a plane of an extended ionic crystal, and a synthetic potential energy model with a stable spacing
. Original schematic geometry and calculated model curves, not a measured electron density, diffraction pattern, or experimental lattice energy. Four nearest neighbours are visible in the plane of the rock salt structure; two more lie perpendicular to it. Disk sizes are arbitrary. The right-hand curves use the explicitly stipulated dimensionless model below. Details.For a large one-dimensional chain of alternating charges, is the Madelung constant. Summing the Coulomb interaction about one ion and dividing by two avoids double counting; per ion pair the result is . There are two nearest-neighbour bonds per pair, so a repulsion per bond gives the displayed second term. If , is a stable minimum with .
A hard-sphere model relating the radius ratio of opposite-charge ions to a possible coordination number. For a cube of eight mutually touching anions around a touching cation, ; real bonding and coordination-dependent radii limit its predictive power.
A geometric heuristic favouring the largest coordination number compatible with neighbouring opposite-charge ions touching the central ion without overlap. A limiting radius ratio rule comes from additionally making the surrounding ions just touch. It is a hard-sphere model, not an exact stability criterion.
A salt is an assembly of cations and anions with zero total electric charge. Its constituents can be single chemical atoms or groups joined internally by covalent bonds. Thus ionic bonding and covalent bonds can occur in the same chemical substance, at different structural levels.[ref]
Magnesium chloride is an ionic compound with two chloride anions per magnesium cation. Complete dissolution supplies one divalent magnesium ion and two monovalent chloride ions per formula unit. At analytical salt concentration , these species contribute molar ionic strength , provided additional dissolved ions are absent.
For a neutral formula unit, let count ions of type with electric charge , where is the elementary charge. The stoichiometry must satisfyFor and , the smallest positive counts are . For and , they are , giving calcium chloride : . The formula counts ions, not the number of nearest neighbours. A counterion is the oppositely charged partner needed to balance a charged species; its identity can change solubility and other properties without changing the identity of that species.[ref]
Historical formulation name; composition is identified by pharmacopoeial edition.
An ionic crystal repeats an arrangement of ions in three dimensions. A unit cell describes that repetition, whereas a formula unit describes composition. The two accounting devices answer different questions. Coulomb's law is not restricted to an arbitrarily drawn line connecting one cation to one anion: every ion contributes to the electric field experienced by the others.
The rock salt structure of ordinary sodium chloride has a coordination number of six for each kind of ion: six nearest neighbours of the opposite sign surround each ion along three perpendicular directions. Its conventional cubic unit cell contains four formula units; sharing corner, face, and edge sites between adjacent cells must be included in that count. Sixfold local coordination is compatible with overall stoichiometry.[ref]
The checkerboard plane in the diagram is a slice through this three-dimensional arrangement. It leaves out the two nearest neighbours above and below the plane. Repeating a complete even-by-even patch gives equal numbers of signs; the displayed odd-by-odd crop has a boundary imbalance and is not a complete neutral ionic crystal or a unit cell. Other salts need not have the rock salt structure or the same coordination number.
In this structural use, the coordination number counts nearest neighbouring constituents. State which neighbours are counted and which ionic crystal structure is being described; it is not an ion's electric charge number.
Eight nearest neighbours at cube corners around a central site; fluorite structure cations have this arrangement.
An ion pair is an associated cation and anion. An isolated ion pair lacks the extended neighbourhood of an ionic crystal. In a chemical solution, ion pairs can coexist with more widely separated ions; their abundance depends on conditions. A neutral gas-phase sodium chloride molecule can be treated as a discrete entity, with its own quantum mechanics and dissociation products. That use of molecule does not describe the constituents of ordinary solid sodium chloride.[ref]
The Feynman Lectures on Physics, Volume II by Richard Feynman and others. §8–3, distinction between vaporized NaCl pairs and separated gaseous ions
[ref]Oppositely charged ions remaining in direct contact without an intervening complete solvent shell. Their relative positions can bias substitution stereochemistry even when bond cleavage precedes nucleophile attack.
The bookkeeping steps and conserve electric charge and describe the electron inventories of separated species. They do not establish a chronological mechanism in which a labeled electron jumps between two pre-existing chemical atoms and becomes a classical object owned exclusively by one of them. In a bound material, quantum mechanics determines the electron density and the total energy.
Ionic bonding and a covalent bond are useful limiting descriptions with a continuum between them. Electronegativity helps anticipate unequal sharing of electron density, but a numerical cutoff does not measure a universal fraction of ionic bonding. A partial atomic charge depends on how the electron density is partitioned; integer formal charges are a bookkeeping convention. Using in a point-charge model therefore does not claim that every partition of the actual electron density assigns exactly those values.[ref]
The Feynman Lectures on Physics, Volume II by Richard Feynman and others. §30–2, “Chemical bonds in crystals”: gradations between covalent and ionic bonding
[ref]Chemistry 2e. §7.2, “Electronegativity and Bond Type”: general guide and exceptions
For two point electric charges in vacuum, Coulomb's law gives the interaction potential energywith zero at infinite separation. Here is the distance between them and is the vacuum permittivity. Opposite signs give negative potential energy and attraction; equal signs give positive potential energy and repulsion. For a finite collection of distinct point electric charges, sum over to count each pair once. A bulk ionic crystal requires the extended lattice sum, including both signs, rather than multiplying one attractive pair by the coordination number.[ref]
The Feynman Lectures on Physics, Volume II by Richard Feynman and others. §8–1, equations (8.1)–(8.3); §8–3, many-neighbour lattice sum
At short distances, overlapping electron density makes the simple point-charge picture inadequate. Quantum mechanics, including the Pauli exclusion principle, matters to the steep repulsive contribution. Pure attraction proportional to would collapse as ; it cannot set an equilibrium spacing on its own. A useful illustrative model isHere scales an assumed fixed arrangement, packages its attractive contribution, and represents short-range repulsion. The constants have units energy times length and energy times length to the power , respectively. This is a model, not a fitted description of a named salt.[ref]
The Feynman Lectures on Physics, Volume II by Richard Feynman and others. §8–3, close-distance repulsion and limits of a purely electrostatic lattice calculation
Differentiating gives and , so the stationary spacing is a stable minimum. For the diagram's stipulated , put and . ThenThe dimensionless curves show how competing interactions select a spacing. Their energy scale and exponent are synthetic; neither is a measured lattice energy or a melting prediction.
The Madelung constant, named for Erwin Madelung, is a dimensionless geometrical factor in the electrostatic binding of an ionic crystal. For a neutral lattice of electric charges with nearest unlike-ion distance , Coulomb's law gives the potential energy per ion pair as . At a positive reference site, , with neighboring electric charges . Both attraction and repulsion enter this lattice sum; summing with a neutral-bulk prescription is essential. Short-range overlap repulsion and other contributions must be added before inferring a stable spacing or a thermodynamic lattice energy.
A lattice energy needs a reaction, a sign convention, and a stated amount. Using the lattice-dissociation enthalpy convention for sodium chloride,The reverse lattice-formation enthalpy is negative with the same magnitude when initial and final states match. Chemistry 2e gives for this separation, per mole of sodium chloride formula units. This is about electronvolts per formula unit, not per individual ion. It concerns separated gaseous ions, rather than neutral chemical atoms, molten salt, or hydrated ions in water.[ref]
Chemistry 2e. §7.5, “Ionic Bond Strength and Lattice Energy”, positive convention, reverse convention and NaCl value
Also distinguish an internal energy change from an enthalpy change: . A static point-charge calculation omits motion and thermal contributions. Reversing a reaction reverses its energy change; replacing internal energy with enthalpy additionally changes the defined quantity. The Born-Haber cycle infers a lattice-formation enthalpy by combining other enthalpies, ionization energies, and electron affinities. These imagined stages are a thermodynamic accounting path, not a claim about the actual sequence of a chemical reaction.[ref]
A Born-Haber cycle uses Hess's law to infer a lattice energy from measurable enthalpy differences and isolated-species data. The motivating difficulty is that separating a mole of an ionic crystal into mutually separated gaseous ions is not a convenient direct calorimetry experiment. Instead, compare two accounting routes between the same elemental reference states and the same solid. Because enthalpy is a state function, the signed differences close exactly when all inputs describe compatible states. These routes do not assert a reaction mechanism or a temporal sequence of reaction intermediates.[ref]
For one mole of sodium chloride formula units, the overall formation reaction isThe stoichiometry requires half a mole of diatomic chlorine molecules, supplying one mole of chlorine chemical atoms. A half coefficient counts an amount of substance; it does not describe half of an individual molecule. Write each auxiliary chemical reaction with its own sign:Here is the sublimation enthalpy, the enthalpy of atomization of chlorine per mole of chemical atoms, and the molar ionization enthalpy represented by the ionization energy input in elementary treatments. If the bond-dissociation energy table gives the compatible molar enthalpy for , use half that number for . Both ions in the last step are gaseous, and no aqueous hydration of ions belongs in that step.
Adding these equations cancels the gaseous chemical atoms, the gaseous ions and the free electron, leaving precisely the overall formation reaction. ThusThe closed traversal returns from the solid to the elemental reference states, adding ; therefore . Notice that the signed attachment term is negative. Substituting a positive spectroscopic electron affinity for would reverse the attachment contribution.
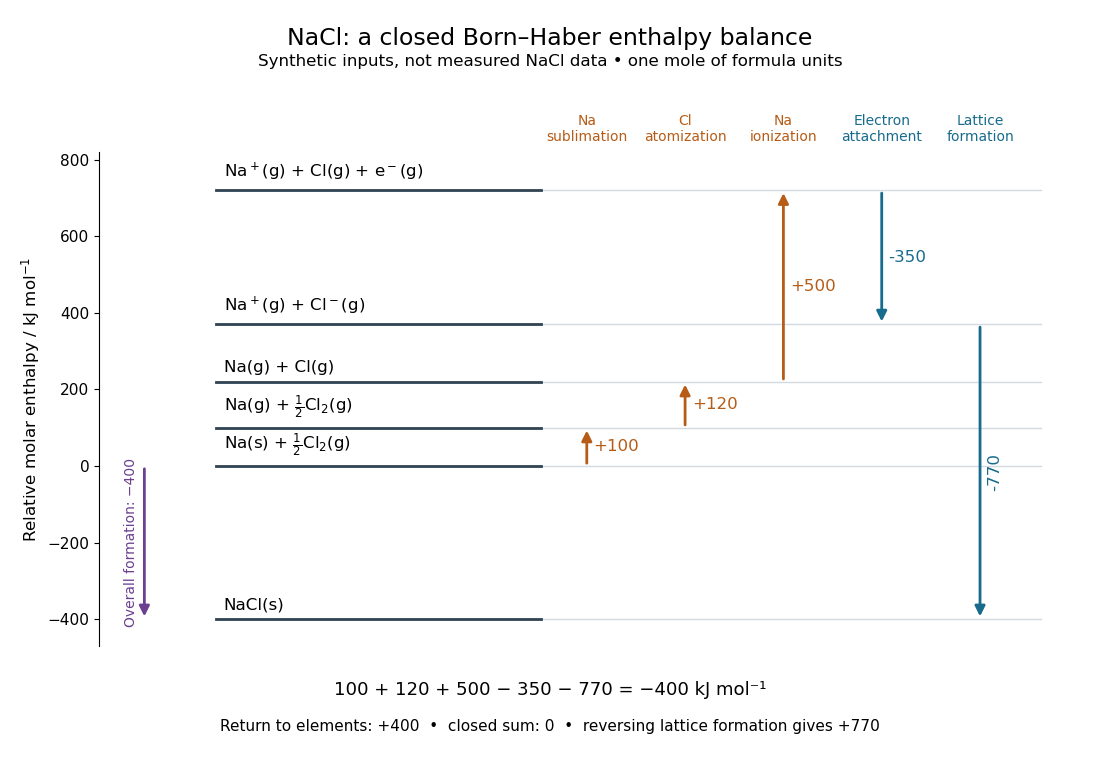
A closed Born-Haber cycle for sodium chloride using explicitly synthetic molar enthalpies: levels represent complete material inventories, and horizontal position is bookkeeping rather than a reaction coordinate
. Details. The diagram deliberately uses invented, internally consistent inputs in : , , , and . Preparation of separated gaseous ions raises the enthalpy by , so and . The closure check is . These numbers are not measured sodium chloride data and do not replace the source's example in lattice energy. Every level retains one sodium nucleus, one chlorine nucleus and zero total electric charge; the ionization level explicitly retains the free electron.
A second indirect route makes the experimental distinction especially clear: a calorimeter can measure dissolution of a weighed salt into a known amount of water, but its final ions are hydrated, not separated gaseous ions. At compatible standard states and the same temperature,Here the combined hydration of ions term takes one mole each of gaseous and to the specified aqueous reference state. Its sum suffices: dissolution calorimetry alone cannot independently assign single-ion hydration of ions enthalpies, or determine without another input. A finite-concentration measurement also requires dilution corrections before it can supply .
In a simple constant-pressure solution calorimeter, an insulated vessel, stirrer and thermometer follow the temperature change after mixing. A calibrated effective heat capacity converts the corrected temperature change to heat. With no applied heating, negligible external heat transfer, and only pressure-volume work, the approximate molar result isHere is the amount of substance dissolved and includes the chemical solution and the vessel. Cooling means positive dissolution enthalpy, because the dissolving sample takes heat from its surroundings.[ref]
Chemistry 2e. §5.2, Figures 5.11–5.13 and discussion of including solution and calorimeter heat capacities
An actual reference measurement used potassium chloride, not sodium chloride: a platinum-lined calorimeter with an adiabatic shield and a quartz-oscillator thermometer. Because dissolution absorbed heat, measured electrical heating prevented loss of shield control; the readouts were electrical input and the vessel/shield temperature record. Stirring was checked visually in a dummy vessel. The thermometer was compared with a calibrated resistance thermometer, and electrical calibration bracketed the reaction. The reported result was at , for one mole of potassium chloride in moles of water.[ref]
The Enthalpy of Solution of SRM 1655 (KCl) in H2O by Marthada V. Kilday. §3 and §4: apparatus, calibration, readouts and specified dilution; inspected through indexed article text
Checks on residual water, particle size and final concentration, and agreement with a second calorimeter, constrained systematic errors. Imperfect shield tracking required heat transfer corrections. This experiment measures an enthalpy of solution, not a direct lattice energy; the hydration and dilution bookkeeping still has to be supplied. The full article could not be opened here; the inspected indexed text supports the stated apparatus and checks, without reconstructing its complete uncertainty budget.[ref]
The Enthalpy of Solution of SRM 1655 (KCl) in H2O by Marthada V. Kilday. §§2–4 and conclusion: moisture, particle-size and dilution studies; confirmation with an isoperibol calorimeter; full-page access blocked
Many ionic crystals have high melting temperatures compared with small molecular solids, because reorganizing their strongly interacting ions costs energy. But melting leaves a dense liquid with interactions; it does not perform the complete separation defining a lattice energy. At fixed pressure, the transition occurs when the Gibbs free energies of the solid and liquid match. The relevant enthalpy and entropy differences both matter, so lattice energy alone is not a universal ordering of melting temperatures. Strong covalent bonds in extended networks can also give high melting temperatures.[ref]
Chemistry 2e. §10.5, “Ionic Solids” and “Covalent Network Solid”; §16.4, equations defining Gibbs free energy and its change
Solubility specifies the amount that can remain dissolved at chemical equilibrium under stated conditions; it is different from how quickly a sample dissolves. Separating ions from an ionic crystal costs energy, but solvation establishes new interactions with the solvent. In water, hydration of ions couples charged ions to polar water molecules. The net enthalpy of dissolution can have either sign; entropy and concentration also enter the Gibbs free energy balance. Consequently, being a salt does not guarantee high solubility in water or any other solvent.[ref]
Chemistry 2e. §11.1, “The Formation of Solutions”, Figure 11.4; §11.3, definition of solubility; §16.4, free energy
Low solubility must also be distinguished from weak electrolyte behavior. A small amount of an ionic compound can dissolve essentially completely as ions, giving few charge carriers because little material dissolved. Calling that a weak electrolyte solely because its chemical solution conducts poorly would conflate two different questions.[ref]
Solvation is stabilization through interactions between a solute and its solvent. It involves both the solute's surroundings and reorganization of the solvent: neighbors move, orientations change, and existing interactions compete with new ones. Electric forces, Van der Waals forces, and specific interactions such as hydrogen bonds can contribute. Solvation can also occur at groups on an insoluble material, so it does not require complete dissolution.[ref]
Solvation by water is called hydration. Hydration of ions is the charged special case, but neutral molecules such as sucrose can also interact with water without becoming ions. A hydrated cation retains its positive net electric charge; orienting neutral water molecules around it does not supply an extra electron. Dissolution, desolvation, and ionization describe different changes, as the examples below show.[ref][ref]
11.2 Electrolytes. Chemistry 2e by Paul Flowers and others; “Ionic Electrolytes” and “Covalent Electrolytes”
The useful question is therefore not just whether a solute attracts a solvent, but how much the whole system gains after paying for the rearrangements. Solvation free energy describes a specified transfer into a solvent; solvation thermodynamic balance explains why even cooling during dissolution can accompany favorable mixing. The existing solubility discussion concerns how much remains dissolved at chemical equilibrium.
A ratio of concentrations or activities between specified phases. State whether all protonation states are combined: a pH-dependent total distribution ratio differs from a partition coefficient for one molecular species.
Reorganization of a solvent to accommodate a solute. Its free-energy cost competes with favourable solute–solvent interactions and depends on solute size and shape.
A solvation shell is the region of solvent constituents surrounding a solute, with local arrangements influenced by their interactions. The first solvation shell contains the closest neighbors under a stated distance or interaction criterion. A coordination number counts those neighbors; it requires such a criterion and can fluctuate. Neither a drawn ring nor a few illustrated molecules determines a measured coordination number.
In a liquid, a solvation shell is an evolving neighborhood: solvent molecules reorient and exchange with the surroundings. Different ions can produce different local dynamics. Spectroscopic measurement of hydration dynamics provides an experimental way to distinguish some of those environments without pretending that a shell is a fixed cage.
A hydration shell is a solvation shell in water. The existing hydration of ions discussion describes preferential orientations around cations and anions.
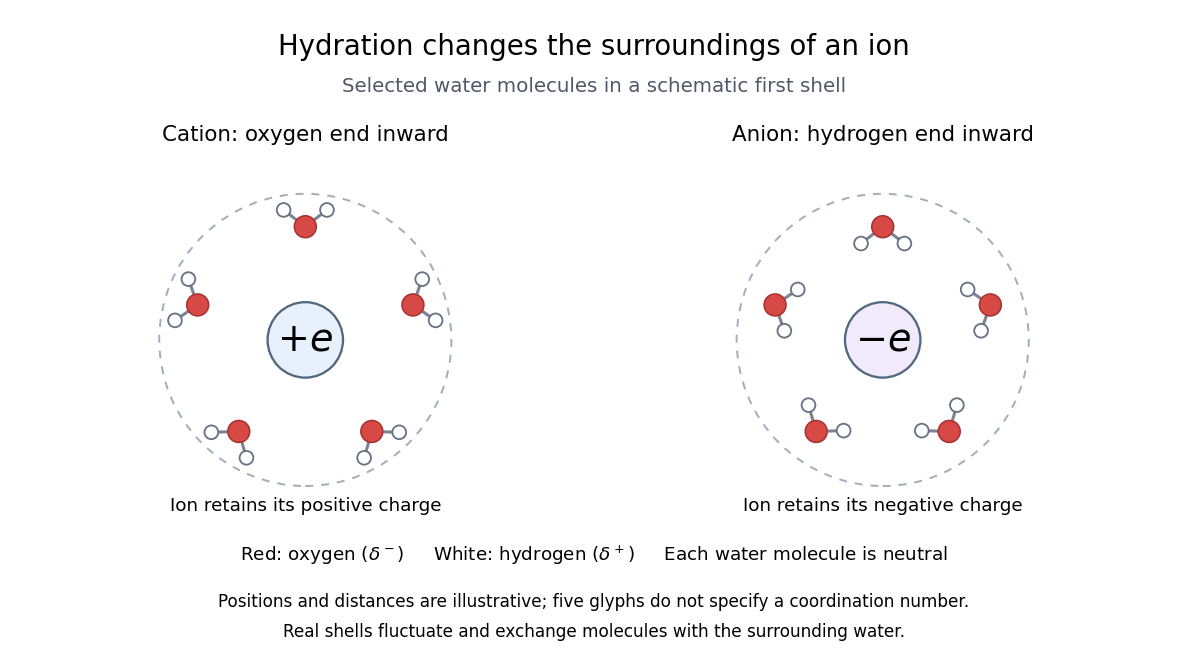
Qualitative hydration shells: opposite orientations of polar water molecules around a cation and an anion, with each ion's net electric charge retained
. Original schematic, not measured molecular positions. Atom spacings and the dashed shell boundary are illustrative. Five selected water glyphs in each panel do not assert a coordination number. Each water molecule is neutral despite its partial atomic charges. Details.An ion-dipole attraction couples an ion's electric field to a molecule's electric dipole moment. In the elementary point-electric dipole approximation, the interaction potential energy is . An orientation with the electric dipole moment along the electric field lowers this energy. For water, the oxygen end preferentially faces a cation, and the hydrogen end an anion. At nonzero temperature, fluctuations and competing hydrogen bonds prevent perfect alignment of every molecule.[ref]
11.2 Electrolytes. Chemistry 2e by Paul Flowers and others; “Ionic Electrolytes”, Figure 11.7 and its explanatory paragraph
What separates a hydration shell from the surrounding water experimentally? Electrical conductivity addresses a different question. The documented lamp demonstration immerses metal electrodes in a chemical solution, rinses them between samples, and records brightness; adding sodium chloride to distilled water makes it conductive. A water blank and dissolved sucrose comparison test whether dissolved material alone explains the signal. Fixed geometry, temperature, and stated concentrations are additional proposed controls. This reveals mobile electric charge, with limits from contamination, electrode interfaces and detection threshold; it supplies no shell positions. Conductivity experiment on salt solutions explains the apparatus and controls in detail.[ref]
Conductivity of Solutions by Harvard Natural Sciences Lecture Demonstrations. “Setting it Up” and “Comments”
In the authored study Influence of ions on the hydrogen-bond structure in liquid water by Anne Willem Omta and others, pump-probe spectroscopy uses tunable mid-infrared pulses traversing a sample. The probe's wave polarization is set at to the pump; after the sample, splitting and polarizers select parallel and perpendicular transmission. Measuring each transmission with and without the pump gives and , the respective logarithms of the transmission ratios. Their dimensionless anisotropy isIts decay probes reorientation rather than merely loss of vibrational excitation. Dilute semiheavy water in heavy water separates the O–H and O–D bands. Varying the semiheavy water concentration checked that transfer of excitation between nearby O–H groups did not determine the decay. These apparatus and control details are from the inspected author-uploaded article, not a new experiment.[ref]
The related thesis chapter documents two O–H absorption bands in a sodium perchlorate chemical solution: approximately for groups forming hydrogen bonds to water, and for groups forming hydrogen bonds to the anion. Selecting pump and probe bands, then a later delay window, separated their responses. The anion-associated anisotropy decayed more slowly; the water-bonded component showed no resolved change in decay time across the tested sodium perchlorate concentrations. The chapter also used O–D excitation to extend observation beyond the shorter O–H vibrational lifetime. These results support distinct local dynamics rather than a permanently fixed hydration shell. Spectral overlap, finite observation windows, and the selected salts limit the inference: this does not prove that every ion leaves all distant water properties unchanged.[ref]
Pump-probe spectroscopy measures a sample's response to a probe pulse at a controlled delay after an exciting pump pulse. Varying delay follows dynamics; selecting frequency and wave polarization can distinguish different contributions. Spectroscopic measurement of hydration dynamics is an application to water surrounding ions.
Articles were limited to the first 100 out of 165 total.
Articles by others on the same topic

A chemical bond is a lasting attraction between atoms, ions, or molecules that enables the formation of chemical compounds. Chemical bonds are fundamental to the structure and properties of substances and are involved in chemical reactions. There are several main types of chemical bonds: 1. **Ionic Bonds**: Formed when one atom donates one or more electrons to another atom, leading to the formation of charged ions.



