Past exam of the chemistry B course of the University of Cambridge 2021 Paper 2 B9 d Solution by  Codex 0 2026-10-20
Ring cleavage of 1,3,5-Trioxane regenerates formaldehyde, whose strong carbon–oxygen pi bond stabilizes the monomer; in water much of it is further stabilized as its hydrate. acetal hydrolysis, particularly under acid catalysis, provides a route between the cyclic trimer and these monomeric forms.
The corresponding formal depolymerization of a siloxane would create silanone units. Such ordinary small-substituent silanones have much less favourable pi bonding and are very reactive toward forming new Si–O–Si linkages. The connected siloxane is consequently preferred over a persistent aqueous solution of silanone monomers. This explains the intended contrast, but aqueous stability is conditional: acid or base can catalyse siloxane cleavage to silanols, and those silanols can condense again. Neither neutral-water inertness of every siloxane nor instantaneous neutral-water dissociation of every trioxane should be inferred.
Past exam of the chemistry B course of the University of Cambridge 2021 Paper 2 B9 c Solution by Codex 0 2026-10-20
Codex 0 2026-10-20boron is sufficiently small for effective oxygen-/boron- overlap, and three-coordinate planar boron benefits from the pi donation just described. A six-membered boroxine ring provides this overlap without severe angle strain. The larger aluminium centre has poorer oxygen-/aluminium- matching and remains strongly Lewis acidic. It can lower its electron deficiency by accepting additional oxygen coordination, making a three-dimensional aluminoxane cage favourable. Four-coordinate aluminium does not require a boroxine-like planar pi system.
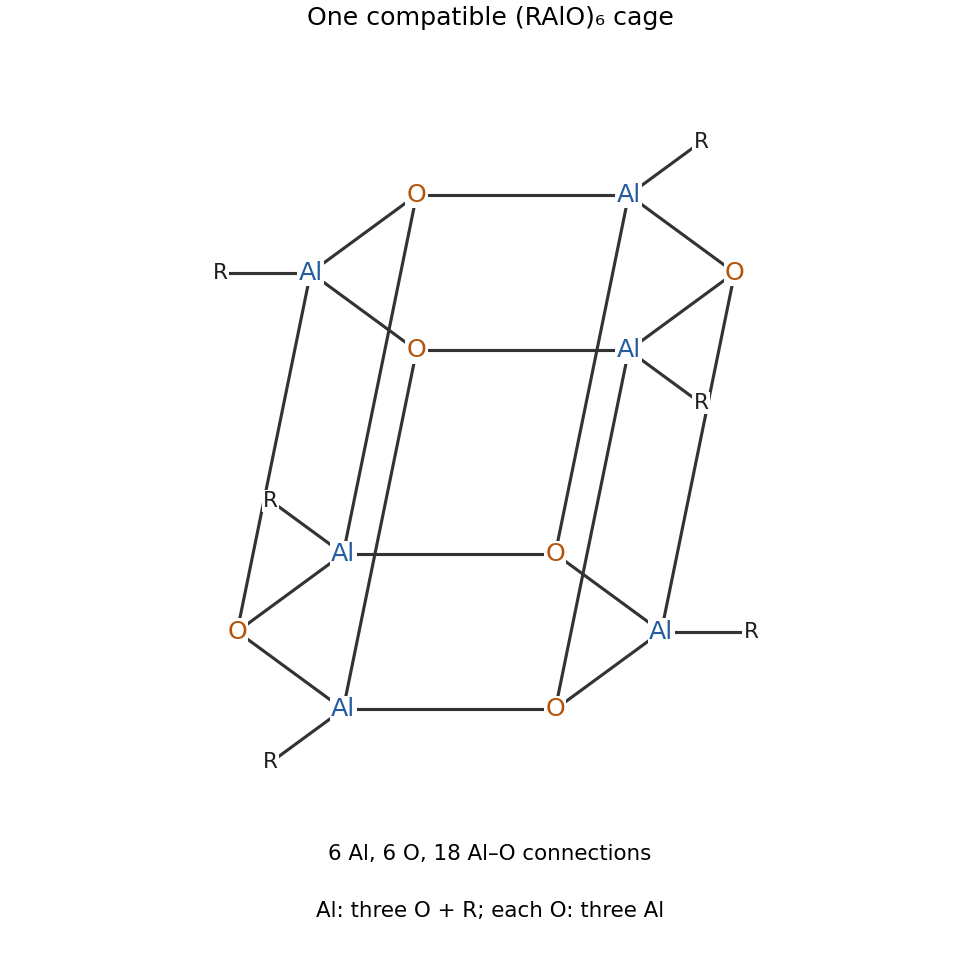
An appropriate hexamer model is an alternating Al/O hexagonal prism, with six aluminium and six oxygen vertices. Colour the twelve-vertex prism alternately Al and O: each aluminium then has three oxygen neighbours and an outward organic substituent, making it four-coordinate, while each oxygen bridges three aluminium centres. The graph has 18 Al–O edges, hence six Al centres with three Al–O bonds each. Different polyhedral connectivities and sizes can meet similar local coordination requirements, so the hexamer is a proposal rather than a claim that the empirical repeat unit fixes a unique cage.
Past exam of the chemistry B course of the University of Cambridge 2021 Paper 2 B9 b Solution by Codex 0 2026-10-20
Codex 0 2026-10-20In a boroxine, each boron has three sigma bonds and is approximately trigonal planar, with an description for its sigma framework and an available perpendicular orbital. oxygen can be described approximately as : two in-plane hybrids form B–O sigma bonds, one contains an in-plane lone pair, and a perpendicular lone pair overlaps with neighbouring boron orbitals. These three oxygen donor pairs supply a six-electron pi system around the alternating ring. The combination of good same-shell overlap and a low-strain six-membered ring favours planarity and partial B–O multiple-bond character. The bonds remain polar; a six-electron diagram alone does not establish benzene-like aromatic stabilization quantitatively.
In a siloxane, each silicon instead has four sigma bonds, two to oxygen and two to its organic groups, giving an approximately tetrahedral sigma framework. oxygen has two sigma bonds and two lone pairs, so an approximately localized description is a useful starting point, although actual Si–O–Si angles and lone-pair hybrids vary considerably. There is no comparable ring of vacant silicon orbitals. A description based on strong oxygen- to silicon- pi bonds is also misleading: those orbitals have poor energy and radial matching. Strongly polar Si–O sigma bonding, supplemented by negative hyperconjugation from oxygen lone pairs into appropriate Si–R antibonding orbitals, gives a better account. The ring can therefore pucker rather than requiring continuous planar pi overlap. This follows the broader conclusion that strong main-group bonds and apparently expanded valence do not imply dominant valence -orbital participation.[ref]
Chemical bonding in hypervalent molecules. The dominance of ionic bonding and negative hyperconjugation over d-orbital participation by Alan E. Reed and Paul von Ragué Schleyer. JACS 112, 1434–1445: ionic contributions and negative hyperconjugation versus d-orbital models
Past exam of the chemistry B course of the University of Cambridge 2021 Paper 2 B9 a Solution by Codex 0 2026-10-20
Codex 0 2026-10-20For the two-atom inorganic backbone unit, and . Their valence-electron sums also agree: . Since each unit bears the same two organic groups, and are isoelectronic species. This electron-count relationship does not require identical bond localization, bond angles or chemical reactivity.
Past exam of the chemistry B course of the University of Cambridge 2021 Paper 2 B8 c Solution by Codex 0 2026-10-20
Codex 0 2026-10-20Use neutral electron counting consistently: each terminal Cp supplies five electrons, while the eight pi electrons of neutral pentalene must be allocated across the shared fused framework rather than counted twice as two independent Cp coordination ligands. One set of 18-electron-consistent proposals is:
- Iron: syn - pentalene with an Fe–Fe single bond. The two iron–Cp fragments lie on the same face. Assigning four pentalene electrons to each iron gives before direct bonding; the Fe–Fe bond contributes one electron to each count, giving 18. Its direct bond naturally accounts for the especially short metal–metal separation. An anti arrangement cannot retain this short same-face bond.
- Cobalt: - pentalene without a Co–Co bond. The count is at each metal. Syn and anti arrangements are possible at this formal level because there is no required direct bond. Removal of the bond relative to iron accounts for a larger separation.
- Nickel: slipped - coordination to the two outer allylic portions. The fusion bond remains an uncoordinated alkene in this limiting bonding picture, while each outer three-carbon portion contributes three electrons in neutral counting: . Equivalent ionic counting gives a nickel(II) centre, a six-electron Cp and a four-electron allyl ligand. Moving the metal binding sites toward the outer three-carbon portions increases the separation further. Syn and anti arrangements can again be drawn.
The original schematic shows the donor regions and the required iron–iron bond; dashed metal-to-region lines denote delocalized coordination rather than localized carbon–metal single bonds.
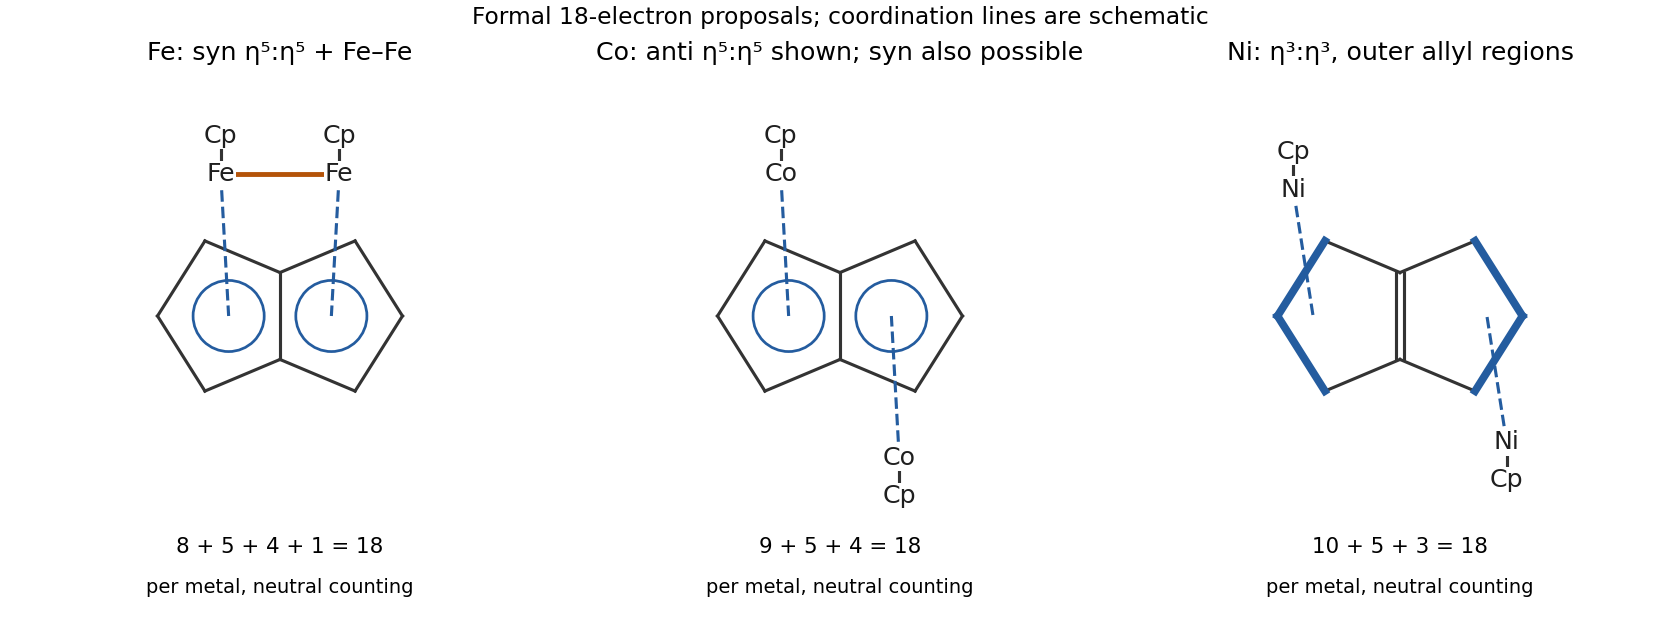
These proposals explain the requested electron counts and distance ordering. electron counting alone does not establish the unique ground-state structure: orbital calculations on related pentalene complexes also identify stable formally electron-deficient arrangements. In particular, the formal requirement of a metal–metal bond in the proposed 18-electron iron model should not be converted into a claim that every known iron pentalene complex adopts that model.[ref]
The Versatility of Pentalene Coordination to Transition Metals: A Density Functional Theory Investigation by S. Bendjaballah and others. Discussion of syn/anti coordination, hapticity and electron-count exceptions
Past exam of the chemistry B course of the University of Cambridge 2021 Paper 2 B8 b Solution by Codex 0 2026-10-20
Codex 0 2026-10-20In a metallocene, several orbitals with substantial metal character are nonbonding molecular orbitals or only weakly bonding or antibonding with respect to the strong Cp–metal framework. Adding or removing a small number of electrons from them can therefore change oxidation state or spin without destroying the principal ring–metal bonds. This allows stable compounds on either side of the 18-electron count: for example, the usual neutral counts for chromocene, ferrocene, cobaltocene and nickelocene are respectively 16, 18, 19 and 20. The 18-electron rule remains a useful reference, but occupation of a weakly antibonding orbital is much less disruptive than occupation of a strongly antibonding orbital. Redox stability also depends on geometry, substituents and reaction conditions, so this is a bonding explanation of the observed flexibility rather than a guarantee of equal stability for all counts.
Past exam of the chemistry B course of the University of Cambridge 2021 Paper 2 B8 a Solution by Codex 0 2026-10-20
Codex 0 2026-10-20Let be normal to the Cp ring through its centre. The five carbon orbitals form symmetry-adapted linear combinations with angular patterns , , and . The all-in-phase combination has sigma symmetry about this axis and can interact with . The pair matches by pi symmetry, while the pair matches by delta symmetry. Rotating within either degenerate pair changes the drawing but not the physical interaction.
For the usual eta-five geometry, the metal interactions are ranked qualitatively pi > delta > sigma. The lobes overlap effectively with the occupied ring combinations. The delta interaction is weaker and involves the higher, initially unoccupied Cp combinations; the interaction with the ring is particularly weak because its axial and equatorial parts contribute little net overlap in this geometry. Thus the statement that a sigma bond is always stronger than a pi bond cannot be transferred indiscriminately to this delocalized ring–metal overlap problem. metal and orbitals also have the symmetry of the coordination ligand combination and contribute to overall bonding, so weak overlap does not mean the entire sigma-symmetry channel is absent.
In ionic electron counting, Cp is with six pi electrons in its three occupied ring combinations, hence a six-electron donor. The same coordination ligand is assigned five electrons in neutral counting.
Past exam of the chemistry B course of the University of Cambridge 2021 Paper 2 B7 e Solution by Codex 0 2026-10-20
Codex 0 2026-10-20Photoinduced CO loss gives a 16-electron fragment. In the ester-substituted complex the pendant ester carbonyl oxygen can reach chromium and capture that vacant site. A suitable structure is , retaining the coordinated arene and adding intramolecular coordination of its ester oxygen. The added oxygen lone-pair donation restores an 18-electron count. The gaseous by-product is carbon monoxide.
heptane is a poor donor, but intramolecular ligand trapping avoids needing a donor solvent. Benzene has no pendant donor. Although the methoxy group has oxygen lone pairs, its directly attached oxygen does not have the same reach and favourable geometry as the projecting ester carbonyl oxygen; possession of a lone pair alone is insufficient. For these two substrates, acetonitrile can trap the unsaturated fragment through nitrogen to give . Without a suitable trap, CO recombination can reverse the initial photochemical dissociation.
Past exam of the chemistry B course of the University of Cambridge 2021 Paper 2 B7 d Solution by Codex 0 2026-10-20
Codex 0 2026-10-20triphenylphosphine is a good sigma donor and a substantially poorer pi acceptor than CO. Replacing CO by this phosphine ligand increases electron density at chromium and reduces competition among carbonyl ligands for pi backbonding. Backbonding into each remaining CO therefore strengthens, their internal bond force constants fall, and both remaining carbonyl stretching frequencies are expected to shift downward relative to the parent tricarbonyl.
The two remaining carbonyl ligands have coupled approximately symmetric and antisymmetric stretches; both are generally infrared active in this geometry. One should not treat the spectral change as simply deleting one of the original two bands. The substituted complex still has valence electrons. No numerical frequency shift follows from electron counting alone.
Past exam of the chemistry B course of the University of Cambridge 2021 Paper 2 B7 c Solution by Codex 0 2026-10-20
Codex 0 2026-10-20The methoxy substituent donates electron density into the coordinated arene by chemical resonance. This increases the electron density available at chromium for pi backbonding to CO, weakening C–O and lowering its stretching frequencies. The ester substituent withdraws electron density from the arene, reducing pi backbonding and raising the carbonyl stretching frequencies. For both observed bands, the methoxy-substituted compound has the lowest carbonyl frequency, the unsubstituted compound is intermediate, and the ester-substituted compound has the highest.
In the harmonic oscillator approximation,so the trend reflects an increasing effective C–O bond force constant rather than a change in the CO reduced mass. For example, comparison of the higher-frequency bands gives . This is only an approximate local-bond interpretation because the observed normal modes couple the three CO oscillators. In the idealized local geometry they span , giving two infrared-active frequencies. A substituted arene can lower the exact molecular symmetry and split the pair, even if that splitting is unresolved experimentally.
Past exam of the chemistry B course of the University of Cambridge 2021 Paper 2 B7 b Solution by Codex 0 2026-10-20
Codex 0 2026-10-20The arene and three carbonyl ligands are neutral, so chromium is in oxidation state zero, with a metal count. The eta-six arene contributes six electrons and each CO contributes two:The methoxy group changes the electronic properties of the arene without changing this formal electron count. The complex is a piano-stool complex, with the coordinated arene as the seat and the three carbonyl ligands as the legs.
Past exam of the chemistry B course of the University of Cambridge 2021 Paper 2 B7 a Solution by Codex 0 2026-10-20
Codex 0 2026-10-20Take the M–C–O direction as . carbon monoxide binds through carbon because its occupied donor orbital has a substantial carbon-end lobe. End-on overlap with an empty metal orbital of sigma symmetry gives CO-to-metal sigma donation. At the same time, occupied metal and orbitals overlap sideways with the two corresponding empty CO orbitals, giving metal-to-CO pi backbonding. Both channels strengthen the metal–carbon bond, while backbonding lowers the internal C–O bond order. The processes are synergic: sigma donation helps supply metal electron density for backbonding, while backbonding relieves electron accumulation at the metal.
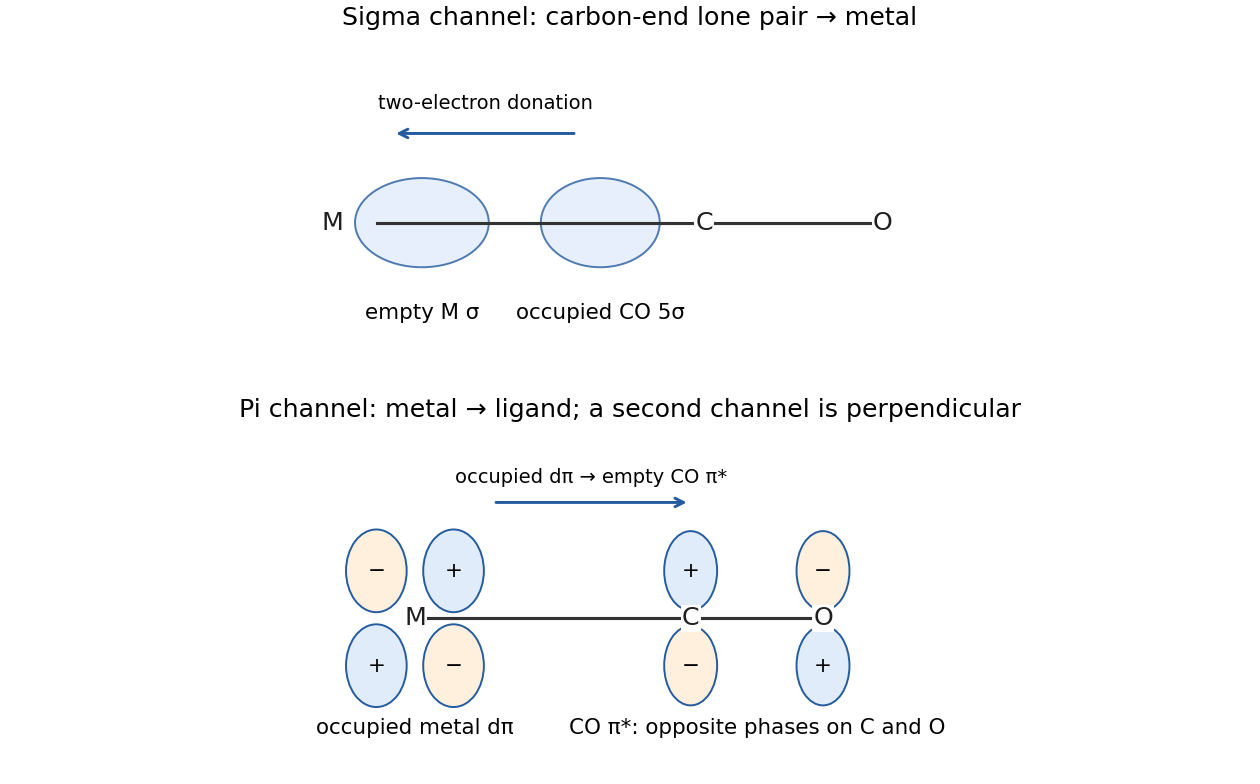


The original orbital sketch shows one of the two perpendicular pi channels. It is the occupied metal orbital, rather than an empty metal orbital, that supplies electrons to CO antibonding orbitals.
Past exam of the chemistry B course of the University of Cambridge 2021 Paper 2 B6 e Solution by Codex 0 2026-10-20
Codex 0 2026-10-20A distinct possibility is terminal sulfur coordination, . Unlike the sulfur donor in the layer, this sulfur need bind only one metal. The limiting resonance structure suggests and . These are single-/triple-bond-based estimates, not exact predictions for an unspecified metal complex. The M–S–C linkage should be bent, consistent with sulfur lone-pair donation.
Past exam of the chemistry B course of the University of Cambridge 2021 Paper 2 B6 d Solution by Codex 0 2026-10-20
Codex 0 2026-10-20For A, linear nitrogen coordination with predominantly nitrile-like bonding is consistent with all three measurements:The short C–N and long C–S distances correspond approximately to a triple bond and a single bond. An approximately nitrogen donor has its donor lone pair directed along the N–C axis, explaining the linear M–N–C angle.
For B, bent nitrogen coordination with increased chemical resonance character accounts for the longer C–N and shorter C–S bonds. In this limiting description both internal bonds have substantial double-bond character and nitrogen is approximately . One nitrogen lone pair can bind a metal with an M–N–C angle near ; alternatively, donation of both nitrogen lone pairs can give a trigonal, nitrogen-bridging unit:The nitrogen-bridging drawing is a compatible proposed bonding mode, but the listed internal distances and one angle alone do not prove the number of attached metals. In particular, B cannot be identified simply as terminal sulfur coordination: the specified angle explicitly contains an M–N bond. resonance structures explain relative bond orders; they do not imply an isolated coordination ligand with permanently localized integer bonds.
There are unlisted articles, also show them or only show them.