Why can two individually fair allele transmissions fail to be independent? Genetic loci on the same chromosome can travel together through meiosis. This tendency is genetic linkage; in an ordinary two-locus model, detectable genetic linkage means a recombination fraction below . A genetic marker can therefore help track a neighboring genetic locus through a pedigree. See Clark, Douglas and Choi, Biology 2e: chromosomal theory and genetic linkage.
Take a diploid individual heterozygous at two genetic loci, with phase-known genotype : one homologous chromosome carries alleles , the other . The parental haplotypes are ; the recombinant haplotypes are . Assume fair Mendelian segregation, equal probabilities within each reciprocal pair of gametes, and no allele-dependent differences in gamete recovery. If the recombination fraction is , the gamete probabilities are in that order. Each genetic locus still transmits its two alleles equally, but exceeds : fairness at each genetic locus does not imply independence between genetic loci.
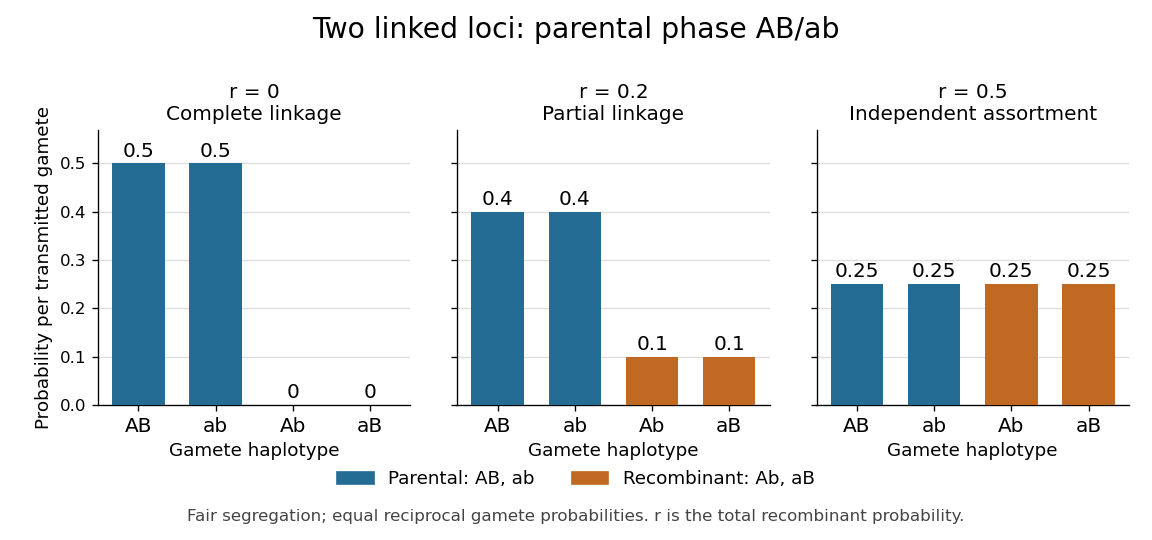
Gamete probabilities from the phase-known genotype AB/ab: parental versus recombinant haplotypes at three recombination fractions
. Details. 
At , only the parental haplotypes occur, each with probability : this is complete genetic linkage. At , all four haplotypes have probability , giving independent assortment. A recombination fraction near alone cannot establish whether the genetic loci lie on different chromosomes or far apart on one chromosome.
Genetic linkage concerns joint transmission through meiosis. Linkage disequilibrium concerns haplotype frequencies in a population, while genetic association compares allele or genotype frequencies with a phenotype. These questions require different observations; pedigrees directly reveal co-transmission.
A genetic map turns observations of inherited haplotypes into an ordered account of genetic loci along a chromosome. Its distances describe genetic recombination, which makes the map useful for locating a genetic locus relative to known genetic markers. A genetic map measures inheritance separation; separation measured in base pairs describes positions along DNA. Recombination hotspots help explain why equal numbers of base pairs need not give equal genetic map distances. See T. A. Brown, Genomes, 2nd edition (2002), chapter 5: Mapping Genomes.
Write for the expected value of the number of chromosomal crossovers encountered along one transmitted chromatid between the chosen genetic loci, measured in morgans; is the distance in centimorgans. This counts exchanges along that transmitted chromatid, rather than all exchanges in the four-chromatid meiotic bundle. For a short interval with very few multiple exchanges, the recombination fraction satisfies : about recombinant gametes then corresponds to about one centimorgan.
For a longer interval, hidden even numbers of chromosomal crossovers make that approximation inaccurate. Under the Haldane mapping function, morgans gives ; conversely gives morgans, or centimorgans. Applying the inverse formula to an already specified genetic map distance would confuse a count with a probability.
Genetic map distances add along ordered adjacent intervals because the total exchange count is the sum of its interval counts. For independent random variables describing counts in disjoint intervals of the Haldane mapping function model, the endpoint recombination fraction instead obeysExactly one interval must change parental origin. Thus gives , although the two genetic map distances add to approximately morgans. The simple composition formula requires the stated independence; crossover interference can invalidate it.
The Haldane mapping function connects additive genetic map distance to endpoint recombination fraction in a toy model with no crossover interference. Specify the model directly along a transmitted chromatid: the exchange count in the interval has a Poisson distribution with expected value , where is the distance in morgans. Counts in disjoint intervals are independent random variables. This is the transmitted-chromatid convention; is not the mean number of exchanges across all four chromatids in meiosis.
An odd exchange count changes parental phase. Summing the odd terms of the Poisson distribution givesFor , the inverse and short-interval approximation areHere is the natural logarithm. At , the recombination fraction is ; at , the distance is morgans, or about centimorgans. The linear approximation is useful only near zero. The recombination fraction approaches as grows; has no finite inverse in this model.
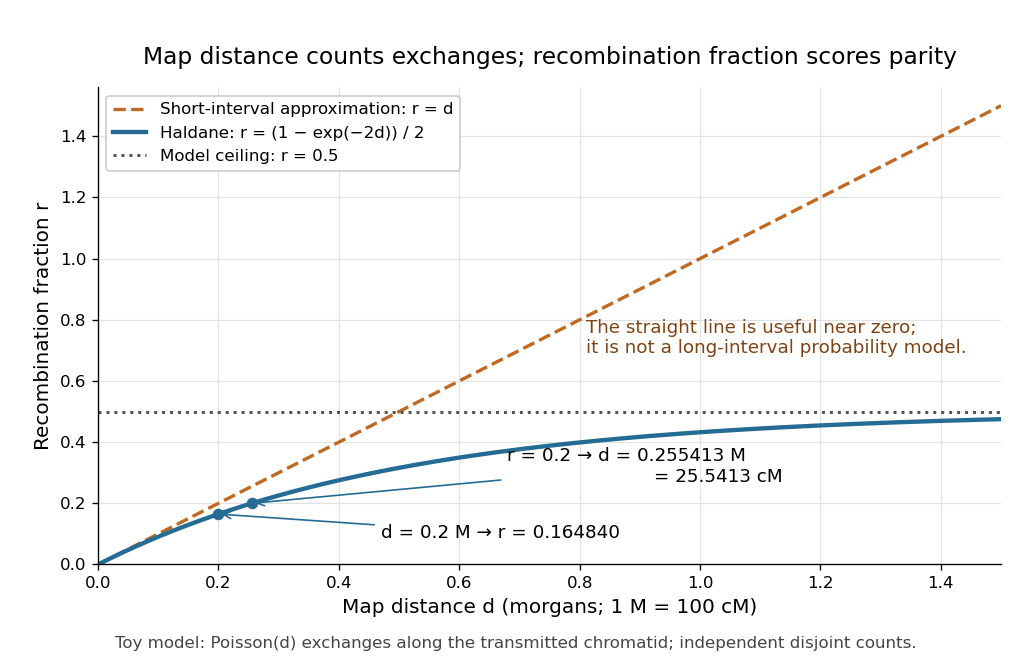
Haldane mapping function versus the short-interval approximation: genetic map distance keeps growing while the recombination fraction approaches one half
. Details. 
The model is a useful baseline, not a universal rule for every interval during meiosis. Crossover interference can change the relationship between distance and recombination fraction. For the count convention and assumptions, see Weeks, Tang and Kwon (2009): genetic map functions and exchange-count conventions. Haldane's original paper is bibliographically recorded as The combination of linkage values, and the calculation of distances between the loci of linked factors, Journal of Genetics 8 (1919), 299–309, in Selected Genetics Papers of J. B. S. Haldane, edited by K. Dronamraju (1990), contents.
A morgan is a unit of genetic map distance: an expected value of one exchange along the transmitted chromatid in the interval. One morgan equals 100 centimorgans; it does not mean a recombination fraction of one.
A centimorgan is one hundredth of a morgan. For a sufficiently short genetic map interval, one centimorgan corresponds approximately to a recombination fraction, rather than an exact identity for all intervals.
Nonparametric linkage analysis tests excess identity by descent among relatives with a shared phenotype, without specifying its complete penetrance model or disease-allele frequency. Its null still uses Mendelian segregation and the specified pedigree.
// Target: biology.bigb
For independent pairs of affected full siblings with unrelated, noninbred parents and a fully informative genetic marker, the no-linkage null gives shared identity by descent count . Therefore and . An excess-sharing test uses the displayed approximately standard normal statistic and an upper-tail critical value. Pairs sharing family members are dependent, so their variance needs separate treatment. This targeted test differs from an omnibus test of all three sharing probabilities.
An affected-sib-pair analysis examines marker identity by descent between affected full siblings. Without genetic linkage to the disease locus, the probabilities of sharing zero, one and two copies are . Excess sharing supplies evidence for linkage under disease models that preferentially transmit a common risk allele to both siblings.
// Target: biology.bigb
For unaffected genetic carriers of a disease with complete penetrance and recessive inheritance, suppose parental marker-disease haplotype phases are equally likely and the marker is fully informative. Write and , where is the recombination fraction. Conditional on sharing a parental marker copy, the probability that both siblings inherit that parent's disease allele is ; conditional on inheriting different marker copies it is . Independent paternal and maternal transmissions therefore give . Conditional instead on both siblings being affected, the IBD probabilities are , where .
// Target: biology.bigb
For independent affected sib pairs with IBD counts , the likelihood function of the recombination fraction is proportional to , with . At fixed sample size, the total shared-copy count is a sufficient statistic by the Fisher-Neyman factorization theorem. Thus a one-sided likelihood-ratio test has the same ordering as a test for excess mean IBD sharing. This equivalence depends on the stated fully penetrant recessive model and does not extend to arbitrary disease models.
// Target: biology.bigb
Parametric linkage analysis forms a pedigree likelihood from a specified disease model, including penetrance, allele frequencies and recessive inheritance or another transmission model. It estimates or tests the recombination fraction between a disease locus and a genetic marker.
// Target: biology.bigb
Complete linkage in a two-locus model means recombination fraction zero: only parental haplotype combinations are transmitted. Multiple genetic markers in a nonrecombining block represent one haplotype segregation observation.
Homozygosity mapping searches for shared homozygous marker regions in individuals with a rare recessive condition, often in an inbred pedigree. Under a single ancestral disease copy, affected individuals inherit two copies identical by descent. Marker homozygosity alone gives identity by state, so founder haplotype frequencies matter. If a particular haplotype has founder frequency and distinct founder chromosome copies underlie all selected descendant copies, their marker-pattern probability is . For complete linkage and a single disease copy, the affected-only numerator is , yielding a LOD score under linkage-equilibrium founder assumptions.
The LOD score compares the likelihood at recombination fraction with independent assortment:When phenotype likelihood is independent of , it can be computed from marker probabilities conditional on phenotype and ascertainment. Positive values support linkage; the observed marker pattern must be specified precisely because affected-only and affected-plus-unaffected observations give different likelihoods.
Articles by others on the same topic
There are currently no matching articles.