Molecular orbital theory describes the electronic arrangement of a molecule using molecular orbitals: one-electron spatial wavefunctions extending over the combined atomic nuclei. Filling bonding molecular orbitals can stabilize a chemical bond; filling antibonding molecular orbitals opposes that stabilization. A molecular orbital is a building block of a many-electron description, rather than the entire many-electron wavefunction. See Robert W. Field, MIT 5.61, lecture 24, pp. 1–4: molecular orbitals and the LCAO approximation.
First make the interference visible with an exactly normalized one-dimensional toy. Replace the three-dimensional atomic orbitals of the hydrogen atom by two Gaussian functions centred at , with dimensionless and :The normalized sum and difference are , following the same linear combination of atomic orbitals algebra. At , the orbital overlap is . At the midpoint, , givingBoth probability densities have total integral one: interference redistributes probability, rather than creating an extra electron. These are toy wavefunctions, not actual hydrogen atom orbitals or predictions of molecular hydrogen energies.
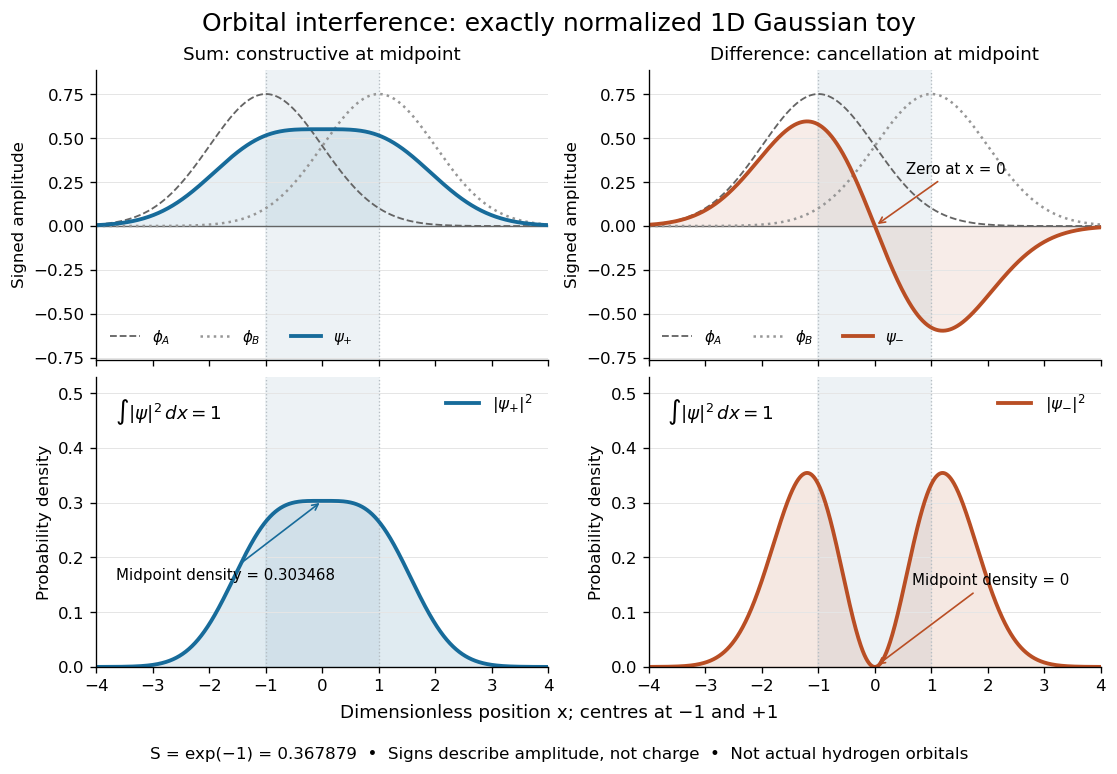
Normalized one-dimensional Gaussian function toy: signed probability amplitudes add or cancel before their squares give probability densities
. Centres at and give orbital overlap . The difference has a zero at the midpoint; its negative lobe does not carry positive electric charge. All density curves shown are normalized on the line, rather than in three-dimensional space. Details.Now fill the two -derived molecular orbitals in the minimal model. The lower bonding molecular orbital is labelled ; the upper antibonding molecular orbital is . The Pauli exclusion principle permits at most two electrons in each spatial molecular orbital, with opposite spin projections along a chosen axis. For the lowest electron configuration in this model, fill first. The subscripts and denote even and odd behaviour under inversion through the midpoint; the star marks an antibonding molecular orbital. See Flowers et al., Chemistry 2e, 8.4: occupancy and bond order and Field, MIT 5.61, lecture 25, pp. 1–3: bonding, antibonding, and symmetry labels.
With electrons in bonding molecular orbitals and in antibonding molecular orbitals, the model's bond order is . The three elementary counts are:
- Molecular hydrogen, : two electrons give and .
- The hydrogen molecular ion : one electron gives and .
- Two neutral helium chemical atoms, : four electrons give and in this minimal model.
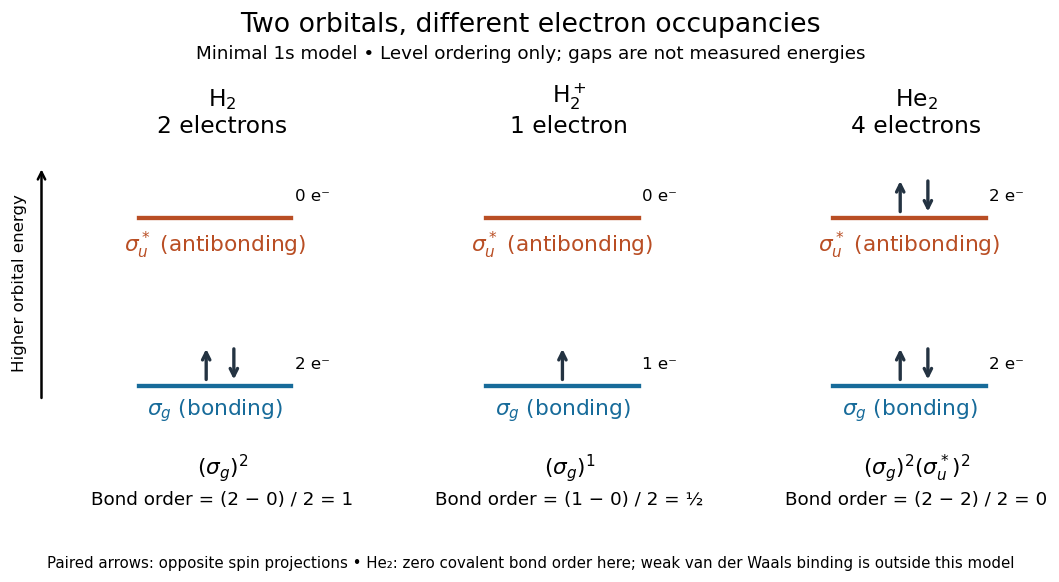
The zero bond order says this minimal model gives no net covalent bond for . It does not rule out every bound pair of helium chemical atoms: a very weak Van der Waals force can bind a neutral helium dimer, which has been observed experimentally. See Zeller et al., Imaging the He2 quantum halo state using a free electron laser, PNAS 113, 14651–14655 (2016), p. 14651: van der Waals binding and Luo et al., The weakest bond: Experimental observation of helium dimer, Journal of Chemical Physics 98, 3564–3567 (1993). The bond order count does not include that weak binding mechanism.
The useful surprise is that sharing alone does not determine a chemical bond: the relative quantum phase and the occupancy matter together. The same two shapes explain why adding an electron to a bonding molecular orbital increases the model's bond order, while adding one to an antibonding molecular orbital decreases it. Molecular orbital theory also accounts for the unpaired electrons and paramagnetism of molecular oxygen, which its all-paired Lewis structure misses. See Chemistry 2e, 8.4: the molecular orbital description of oxygen.
A correspondence between molecular fragments based on the number, approximate symmetry, shape, energy and occupation of frontier orbitals. Equal total electron count alone is insufficient; the comparison predicts transferable bonding possibilities rather than identical chemistry.
A diagram linking reactant and product orbitals by a symmetry conserved along a stipulated reaction path. Orbitals of the same symmetry avoid crossing in a generic one-parameter problem; orbitals of different symmetry can cross. Correlation constrains connectivity, not quantitative barrier heights.
Mixing between symmetry-compatible s-derived and p-derived orbital combinations. It shifts energies and changes orbital shapes; in a centrosymmetric diatomic, gerade and ungerade combinations do not mix with one another.
A model of conjugated molecular orbitals with an orthonormal atomic basis, one-centre energies , and resonance matrix elements between selected neighbouring sites. Solving gives orbital energies and coefficients. Its simplicity makes symmetry and electron filling explicit, but quantitative charge predictions can depend strongly on the chosen parameters and omitted orbital relaxation.
With one orthonormal site orbital per cube vertex, common site energy , and coupling along every edge, the one-electron Hamiltonian is , where is the cube adjacency matrix. Its eigenvalues are with multiplicities .
One proof labels each vertex by three signs. The adjacency operator flips one sign at a time, so a product of coordinate signs is an eigenfunction of with eigenvalue . There are such products. For , eight electrons fill the lowest four spatial molecular orbitals, yielding total one-electron energy .
For a four-site cycle with site energy , three resonance integrals and one weakened integral , , put . In the independent-electron Hückel method, filling the two or four lowest spin orbitals gives total electronic energies and . At the square limit , these become and . The weakened-edge distortion raises the two-electron sum but lowers the four-electron sum. Near the latter change is , so its first-order term cancels. This electronic comparison omits structural repulsion and other restoring forces, and does not determine the open-shell spin at the square limit.
For a bipartite pi system with a common diagonal energy , the orbital spectrum is paired about . If and changes the sign on one sublattice, then ; an eigenvector of energy is mapped to one of energy . Their site-probability magnitudes are equal.
For real orthonormal site functions, , where is orbital occupation. It measures the off-diagonal pi density matrix; a complete C–C bond also contains a sigma contribution.
The diagonal one-centre energy assigned to a pi-basis orbital. This one-electron parameter is distinct from a two-electron Coulomb repulsion integral in Hartree-Fock theory.
An off-diagonal Hamiltonian matrix element between selected neighbouring pi-basis functions. The usual bonding convention has ; non-neighbour matrix elements are neglected.
With equal on-site energy and equal pair interaction , the three-site Hamiltonian has one eigenvalue and a degenerate pair . This model explains symmetry and filling but omits ingredients needed for a quantitative dissociation energy.
The change in energy on allowing a specified localized orbital reference to delocalize across a conjugated system. In a Hückel method calculation, the reference must use the same one-centre parameters and bond types as the delocalized molecule. With the signed convention , stabilization is negative; its positive magnitude is often also called the delocalization energy, so the sign convention must be stated.
In an orthonormal Hückel method basis, an occupied molecular orbital with occupancy contributes electrons to site . Adding one electron to a fixed orbital changes the site population by . A coefficient sign indicates orbital phase, not charge sign. A net atomic charge also needs a reference population and treatment of the electrons outside the model.
A molecular orbital is a one-electron spatial wavefunction for the combined atomic nuclei of a molecule. For a normalized molecular orbital, the Born rule gives the position probability density , with . The molecular orbital is the amplitude, rather than its squared modulus or a classical trajectory.
In the independent-orbital approximation, a doubly occupied molecular orbital contributes to the electron number density, whose integral is two. An empty molecular orbital contributes zero, even though its shape is still part of the model. The molecular hydrogen example therefore uses one occupied spatial molecular orbital for two electrons, distinguished by their spins. A negative real probability amplitude means a relative quantum phase of , not positive electric charge; multiplying the whole wavefunction by changes only its global phase.
A molecular orbital near the boundary between occupied and unoccupied levels. The highest occupied molecular orbital and LUMO are central examples. Their energies, occupancies, spatial form and symmetry help predict which new chemical bonds can form when fragments approach.
A molecular orbital whose occupation changes bonding between the selected centres relatively little. In a real complex, weak bonding or antibonding contributions may replace exact nonbonding character.
For an open-shell species, distinguish a singly occupied spatial orbital from a vacant spin orbital. Adding an electron to an oxygen molecule can fill the opposite-spin vacancy in a pi-star singly occupied molecular orbital even though the next completely empty spatial orbital is sigma-star.
A spatial molecular orbital occupied by one electron. In an open-shell molecule, its empty opposite-spin slot can accept another electron even though the spatial orbital is occupied.
A molecular orbital with little or no net bonding interaction between the atoms in the stated orbital model. A symmetry-isolated atomic orbital can be exactly nonbonding in a minimal basis: the out-of-plane nitrogen orbital in planar Ammonia has no matching hydrogen symmetry orbital. A mainly localized lone pair can acquire bonding admixture when a distortion permits additional mixing.
An energy-level scheme showing the occupied molecular orbitals and their relation to atomic orbitals. Occupancies must account for degeneracy and spin.
In pyramidal Ammonia, the hydrogen symmetry orbitals span of the C3v molecular point group. Nitrogen and both have symmetry and span . In the planar D3h molecular point group, these become , and , respectively, while the hydrogen combinations span . Thus the planar out-of-plane lone pair is a symmetry-isolated nonbonding orbital; pyramidalization permits mixing with the other functions.[ref]
The occupied molecular orbital of greatest energy in a specified reference electron configuration. A fixed-orbital model removes an electron from this level first; relaxation and electron correlation can modify the resulting energy.
Orbital degeneracy means distinct spatial orbitals have equal energy within the specified model or symmetry. Two electrons in degenerate orbitals need not pair; exchange can favor a triplet state.
The lowest-energy unoccupied orbital of a specified reference electron configuration. In a fixed-orbital approximation, adding one electron occupies this orbital; orbital relaxation and electron correlation can alter that picture.
Occupying a bonding molecular orbital favours a chemical bond. For the two equivalent atomic orbitals of the hydrogen atom, the sum has constructive interference between the protons. The resulting electron density in that region helps stabilize the combined system. Its lower energy in this model follows from the Quantum Hamiltonian, not from the plus sign alone; see Field, MIT 5.61, lecture 25, pp. 1–2.
For example, in molecular hydrogen supplies , hence bond order one when . The sum in the molecular orbital theory toy has midpoint probability density , whereas the difference has zero there. The normalization factors must be included in this comparison.
Occupying an antibonding molecular orbital opposes a chemical bond. For equivalent atomic orbitals of the hydrogen atom, the difference vanishes on the plane halfway between the protons: every point there is equally far from both centres, so the amplitudes cancel. This nodal plane removes the contribution of this molecular orbital to electron density on that plane; it does not make an entire finite region empty. The -derived antibonding molecular orbital lies above its bonding molecular orbital partner in energy. See Field, MIT 5.61, lecture 25, pp. 1–2.
In the minimal model, adds and cancels the bond order contribution of . Opposite signs on the two lobes describe probability amplitudes, while both lobes have nonnegative probability density after squaring. The star is an orbital label, not an extra electron or an electric charge.
The LCAO approximation builds a molecular orbital as a linear combination of single-centre atomic orbitals. For two equivalent normalized real functions and , choose their signs positive and define their orbital overlap . At nonzero separation, , and the normalized combinations areHere and . Equal coefficients use the equivalence of the two centres; a general LCAO need not have equal coefficients. See Field, MIT 5.61, lecture 24, pp. 3–6: LCAO coefficients and overlap integrals.
For the Gaussian function toy's , the two denominators are and . To check wavefunction normalization, expand the square: . Dividing by gives one. The local probability density isThe cross term is the interference; adding the separate densities first would lose it. These formulas normalize the chosen linear combination exactly, while the restricted choice of atomic orbitals remains an approximation to an actual molecular orbital.
A linear combination of atomic orbitals chosen to transform under an irreducible representation of the molecular point group. Relative orbital signs must be transformed as well as the positions of their centres. Such combinations organize a Hamiltonian operator into symmetry blocks; combinations of the same symmetry generally still mix and need not themselves be molecular orbitals.
Four equivalent atomic orbitals at the vertices of a regular tetrahedron carry the tetrahedral symmetry representation . The normalized equal sum is ; three coordinate-sign combinations span . Every normalized coefficient vector has entries in a suitable orthonormal real basis. These functions simplify a Hamiltonian invariant under the tetrahedral operations, but equivalent symmetry copies on other sites may still mix.
The orbital overlap of normalized real atomic orbitals is their inner product . It is a dimensionless measure of their common amplitude, not an electron count. For the Gaussian function toy, increasing from to reduces from to . As tends to zero, both LCAO normalization denominators tend to .
In the elementary molecular orbital theory count, bond order is half the excess of electrons in bonding molecular orbitals over those in antibonding molecular orbitals. Thus adding one electron to a bonding molecular orbital raises by , while adding one to an antibonding molecular orbital lowers it by . The configurations drawn above give , , and . This count summarizes net covalent bond occupancy; it is neither a measured energy nor a universal test for every kind of binding. See Chemistry 2e, 8.4: molecular orbital bond order.
Articles by others on the same topic



Molecular Orbital (MO) Theory is a fundamental theoretical framework in chemistry that describes the electronic structure of molecules by considering the combination of atomic orbitals to form molecular orbitals. Unlike Valence Bond (VB) Theory, which emphasizes localized bonds between pairs of atoms, MO Theory provides a more delocalized view of electrons in a molecule.