charge stabilization of colloids supplies repulsion when like-charged surfaces and their electric double layers approach. steric stabilization uses attached or adsorbed molecular layers whose overlap costs conformational and osmotic free energy. Combining both gives electrosteric stabilization; a solvated coating must remain attached and well solvated. A viscous medium slows encounters but does not by itself create an equilibrium repulsive barrier.
DLVO theory adds screened electrostatic repulsion to Van der Waals force. A representative close-gap form isfor equal spheres of radius , with Hamaker constant and inverse Debye–Hückel screening length . The formula is schematic near molecular contact, where hydration, surface roughness and finite ion size matter. A repulsive barrier many high produces kinetic stability even if the primary attractive minimum is deeper than the dispersed state. Entry into that primary minimum generally gives strongly bound aggregation; a shallow secondary minimum can give reversible flocculation.
Adding electrolyte increases , shortens the repulsion and can remove the barrier, enabling fast diffusion-limited aggregation. The critical coagulation concentration is the approximate salt concentration at which this rapid-coagulation regime begins under specified conditions. It is not a universal property of the salt alone. Test the prediction using time-resolved dynamic light scattering or turbidity at controlled particle concentration, pH and temperature; check that added ions do not specifically adsorb or change the surface chemistry.
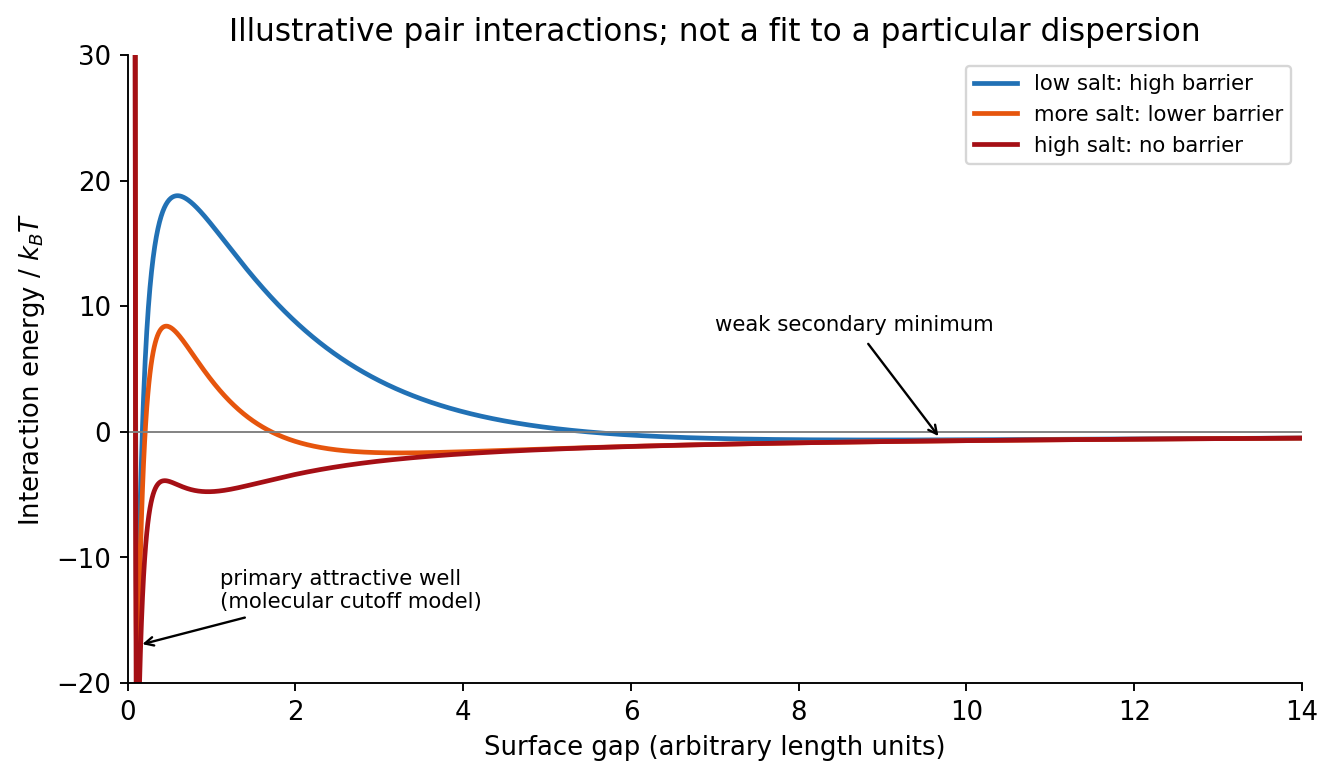
Original schematic DLVO interaction curves showing a stabilizing barrier, secondary minimum and barrier loss on salt addition
. Details. 
In the high-surface-potential Schulze-Hardy rule, the critical coagulation concentration scales approximately as with the counterion valence. HenceThe more highly charged counterion therefore requires about eleven times less concentration in this idealized estimate. This assumes the cations are the counterions of a negatively charged surface, the surface-potential regime is appropriate, and the only changing mechanism is electrostatic stabilization. Specific adsorption and hydrolysis of aluminium ions can invalidate this scaling; matching ionic strengths instead is not the same prediction.
The more negative standard reduction potential identifies iron as the species more readily oxidized. Combining with gives the spontaneous standard galvanic cell reactionThe electron balance is two per reaction, and the activities of the pure metals are unity.
For a galvanic cell, subtract the standard anode reduction potential from the standard cathode reduction potential:A reduction potential is not multiplied by the number of transferred electrons.
Iron is the negative anode and tin is the positive cathode of this spontaneous galvanic cell. electrons travel through the external circuit from iron to tin. An intact tin coating protects iron mainly by physically excluding the corrosive medium. If a defect exposes iron while the metals remain electrically connected, tin is not a sacrificial coating: the exposed iron can become the anodic corrosion site. A small exposed iron area coupled to a large cathodic tin area can be especially damaging; this distinction follows from the electrode-potential order rather than the appearance of the coating.
With , the standard reaction Gibbs free energy and dimensionless equilibrium constant areFor the reaction as written, , and the Nernst equation gives . Consequently zero cell potential requires under the specified unit-activity-coefficient assumption. Reversing that ratio reverses the conclusion. This mathematical ratio does not by itself guarantee experimentally attainable concentrations without precipitation or other chemistry.
A Ragone plot has specific energy on its horizontal logarithmic axis and specific power on its vertical logarithmic axis, normally in and . The comparison must use the same mass convention: complete devices cannot fairly be compared with electrode-active-material values. At approximately constant delivered power, the discharge time isThus constant-time lines slope upward on the log–log plot. Ordinary capacitors lie toward high power and low energy; batteries toward higher energy and lower power; supercapacitors bridge the two. Technology and operating limits produce overlapping regions, not universal sharp boxes.
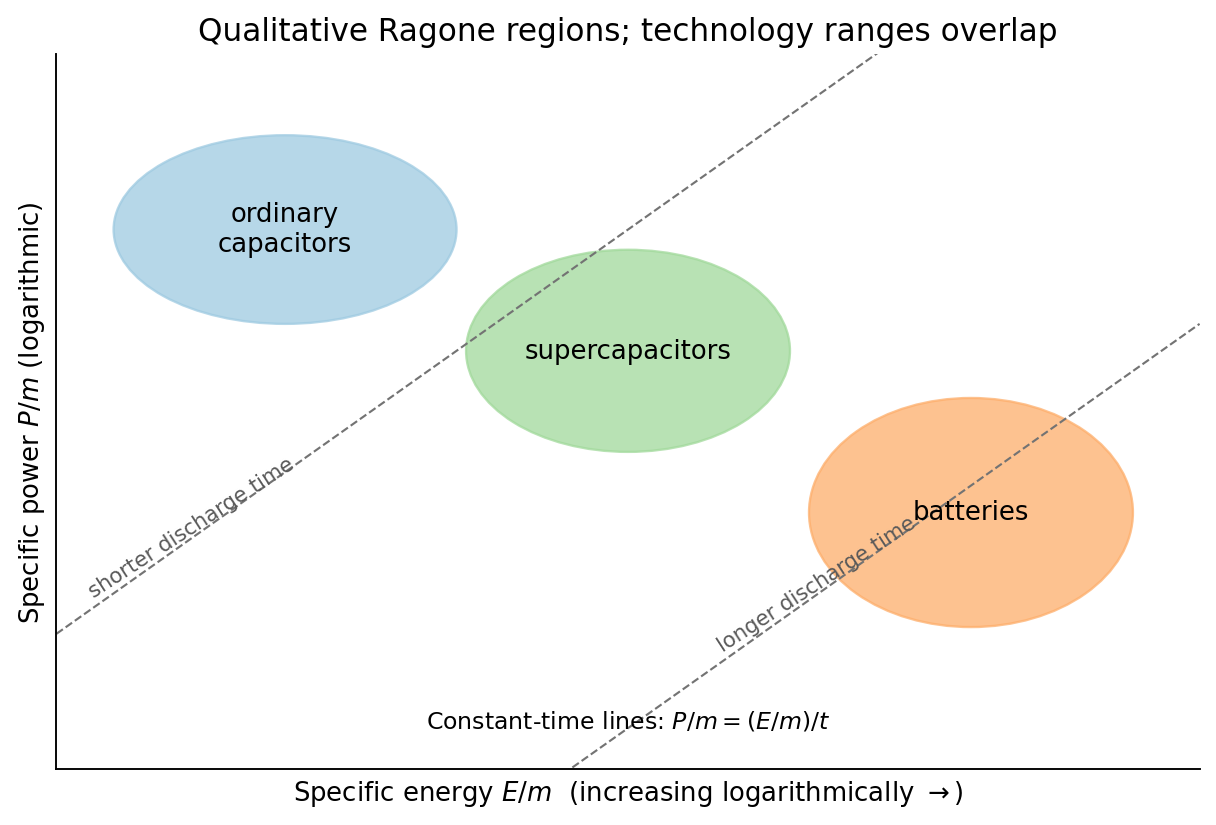
Original qualitative Ragone comparison with logarithmic axes and representative constant-discharge-time lines
. Details. 
An ordinary capacitor stores energy by separating charge across a dielectric: and . Electronic charge redistribution is rapid, giving high specific power, but the dielectric breakdown field and modest capacitance limit specific energy. Series resistance limits pulse power and produces heating; a large nominal capacitance does not remove those losses.
A supercapacitor has a much larger interfacial area and nanometre-scale charge separation. An electric double-layer capacitor stores charge without a bulk redox transformation; pseudocapacitance adds sufficiently fast surface or near-surface redox storage. ion motion in pores, electrolyte stability and series resistance constrain its response. Its specific energy usually exceeds an ordinary capacitor's while high specific power and cycling durability remain possible.
An electric battery stores chemical free energy through coupled electrode oxidation-reduction reactions. It can store substantial energy per mass, but charge transfer, ionic transport, diffusion and heat removal constrain rapid discharge. The terminal voltage and accessible capacity depend on discharge rate and state of charge. Accordingly the plotted order follows the storage and transport mechanisms; it is not a claim that every battery has less power than every supercapacitor.
Articles by others on the same topic
There are currently no matching articles.