Two-dimensional infrared spectroscopy measures a sample's vibrational response as a function of two frequency coordinates. In a coherent experiment, timed infrared pulses establish an excitation history and a later optical response; the spectrum correlates excitation and detection frequencies. The two dimensions are spectral coordinates, not two spatial directions through the sample.
In a time-domain implementation, the experiment scans a coherence delay and resolves a detection interval while holding a 2D IR waiting time fixed. Schematically, a complex measured response is transformed toThe signs and combination of measured response pathways depend on the experimental convention. Fourier analysis converts the scanned delays into frequency information; it does not by itself establish which molecular mechanism produced a feature.
An original spectral example has modes near and . Features at coordinates and lie on the diagonal. An off-diagonal feature at is a 2D IR cross peak: excitation associated with one coordinate is correlated with response at another. Its physical interpretation depends on the pulse sequence and waiting time. A cross peak is not automatically evidence for chemical conversion between two species.
A primary2000 peptide experiment used phase-controlled femtosecond pulses, phase matching and heterodyne detection to measure the phase and amplitude of vibrational photon echoes as pulse delays varied. Cross peaks in acyl-proline-NH spectra supplied evidence for coupling between amide-I modes, with solvent-sensitive structural differences.[ref]
Compared with a one-dimensional infrared spectroscopy trace, the extra coordinate can separate overlapping responses and expose relationships between vibrations. Varying the 2D IR waiting time adds a controlled time dependence to test candidate mechanisms. Interpretation still requires a response model, sufficient time/frequency resolution and controls for solvent and instrument contributions; a richly structured spectrum does not uniquely determine a molecular structure by inspection.
Spectral diffusion is the time-dependent wandering of a transition frequency as its local environment changes. A collection of molecules can retain the same instantaneous frequency distribution while individual members lose memory of their earlier frequencies. In the infrared spectroscopy setting, vibrational spectral diffusion concerns vibrational transitions. The name describes frequency evolution, not necessarily spatial diffusion of the molecules.
A useful original stationary model writes and defines the autocovariance functionFor a real stationary process, its frequency-increment variance isThis follows by expanding the square: stationarity makes the two squared-frequency terms equal, while the cross term is . The relation supplies a spectral-diffusion frequency-increment variance without assuming that the mean transition frequency shifts.
Take the stipulated model , with and . After 2 ps, normalized memory is and the root-mean-square frequency change is . The long-time increment spread approaches , while the single-time spread remains . Thus loss of temporal correlation need not broaden the stationary ensemble distribution over time.
A static distribution offers a contrasting physical limit. Molecules may have different, but individually fixed, frequencies. Then and the increment variance is zero despite a broad ensemble spectrum. A single spectral line width cannot determine how much of that diversity is static or dynamically sampled.
In two-dimensional infrared spectroscopy, controlled infrared pulses first label frequency and later interrogate the response after a 2D IR waiting time. Repeating over waiting times tests the frequency relation rather than merely measuring one ensemble average.[ref]
Frequency-frequency correlation functions and apodization in two-dimensional infrared vibrational echo spectroscopy: A new approach by Kyungwon Kwak and others. Introduction, pulse timing, initial-frequency labeling and later frequency readout
Its calculated spectra illustrate the associated evolution of spectral geometry.[ref]Frequency-frequency correlation functions and apodization in two-dimensional infrared vibrational echo spectroscopy: A new approach by Kyungwon Kwak and others. Fig. 1, calculated spectra and center lines at different waiting times
Environmental fluctuations, population loss and exchange between resolved species affect spectra in different ways. A changing 2D IR line-shape correlation is useful evidence only with an appropriate response model and controls. A fitted time scale is not automatically a molecular transport coefficient or the lifetime of every environmental structure.
The spectral-diffusion frequency-increment variance quantifies how much a transition frequency changes between two observation times. For a real stationary process with mean and autocovariance function ,The mean increment is zero under these assumptions, so its second moment is also its variance. Expanding the square proves the identity: the two squared-frequency deviations each contribute and their cross term contributes .
If , thenFor an original example with and , and the root-mean-square increment is about . The single-time frequency spread remains ; temporal decorrelation does not require the stationary distribution to broaden.
The long-time value assumes that the autocovariance function tends to zero. It is not a universal upper bound. For a random-phase sinusoid with phase uniform over a full cycle,At half a cycle, the frequencies are anticorrelated and ; at a complete cycle the increment vanishes. Thus a nonmonotone increment statistic can reflect recurring frequency memory.
For nonstationary observations , the general identity instead isDrift of the mean must not be silently assigned to stationary spectral diffusion. Likewise, under an explicit model of independent zero-mean endpoint measurement errors with variance , the observed second moment gains . This spectral frequency-increment measurement noise term changes when the endpoint errors are correlated.
Primary two-dimensional infrared spectroscopy work uses controlled frequency labeling and later readout to constrain frequency memory.[ref]
Frequency-frequency correlation functions and apodization in two-dimensional infrared vibrational echo spectroscopy: A new approach by Kyungwon Kwak and others. Introduction, pulse delays and frequency-frequency correlation function
The identity above is a model relation; a measured 2D IR center-line slope needs the stated response assumptions before it supplies .Spectral frequency-increment measurement noise is the contribution of frequency-estimation errors to a statistic comparing a transition frequency at two times. It can create an apparent nonzero spectral-diffusion frequency-increment variance even when the underlying frequency has not changed.
For an explicit measurement model, write and define , . With zero-mean endpoint errors,The last term vanishes only under an appropriate lack-of-correlation assumption between the physical increment and its estimation errors. Merely naming the errors noise does not establish that assumption.
If that cross term vanishes, equal endpoint standard uncertainties and error correlation coefficient giveIndependent errors give . A perfectly shared offset cancels from the difference, although the absolute frequencies can remain uncertain. This correlated spectral endpoint error is an ordinary application of correlated-input uncertainty propagation.[ref]
An original example has true root-mean-square increment and . Independent endpoint errors give and apparent root-mean-square increment . At , the added term is only , giving an apparent increment about .
A noise subtraction can also be uncertain. Observed second moment minus estimated noise contribution gives in the same squared-frequency units. That negative estimate does not establish a negative physical variance; it flags uncertainty or a mismatch in the assumptions used to separate the contributions.
Repeated observations of a stable reference can test the estimator's noise and temporal correlation. Filtering, common references and reuse of the same data can correlate errors at nearby times. Primary two-dimensional infrared spectroscopy analysis connects spectral geometry to a frequency-correlation model rather than directly recording every molecule's frequency trajectory.[ref]
Frequency-frequency correlation functions and apodization in two-dimensional infrared vibrational echo spectroscopy: A new approach by Kyungwon Kwak and others. Introduction and Section II, spectroscopic observables and frequency-frequency correlation function
The additive-error equations apply only when the stated frequency estimator and error model describe the actual measurement.Spectral common-mode offset rejection is cancellation of an additive frequency-axis shift when the same shift enters two spectral estimates with equal sensitivity and the estimates are subtracted. It improves a frequency difference without establishing accurate absolute frequencies. The relevant measurement model must identify what is shared, and over which observations.
For true frequencies , write measured estimates asThe unknown common offset disappears exactly fromFor independent residual errors of standard uncertainty , the difference still has standard uncertainty . Cancellation of is not cancellation of all measurement noise. With correlated residuals, the covariance term must be retained in uncertainty propagation.[ref]
Evaluation of measurement data — Guide to the expression of uncertainty in measurement by JCGM. Section 5.2, correlated input quantities
An original example uses two spectral positions at and . An additive shift of gives measured positions and , with the same separation. A shared gain error is different: gives separation . Subtraction rejects the offset but retains the scale error.
The algebra generalizes to a contrast of several spectral positions. If and each contains the same additive , its contribution is . Thus weights satisfying reject that offset. Averaging positions with weights summing to one does not. This is an original linear-model derivation of the rejection condition, not a claim that every spectral estimator has this form.
Two failures of the ideal assumption are especially informative. With unequal spectral offset sensitivity, , subtraction leaves . If the offset drifts between observations, it instead leaves . For a stationary drift with variance and correlation at lag , this residual has varianceA common instrument or reference does not alone imply perfect cancellation at different times.
An apparatus-level check measures two stable reference features using the same acquisition and fitting pipeline, then deliberately shifts the assigned axis origin. Their separation should stay fixed under an additive reassignment; a scale change should alter it. Repeated reference observations test temporal drift and residual covariance. In two-dimensional infrared spectroscopy, this test concerns frequency-estimation errors; spectral diffusion also describes physical fluctuations and cannot be identified or removed merely by subtracting an instrumental offset.
Unequal spectral offset sensitivity occurs when a shared instrumental or reference offset influences two spectral estimates by different amounts. It limits spectral common-mode offset rejection: subtraction cancels a common disturbance only to the extent that its two response coefficients match.
For frequencies , write an explicit measurement modelwhere is the shared offset, are dimensionless sensitivities and are residual errors. ThenThus even perfectly shared leaves a residual when .
An original uncertainty example uses , and standard uncertainty . The offset contribution to the difference is . With independent residual standard uncertainties of each and no correlation with , the combined uncertainty isA general calculation retains the relevant covariances; the independent-error simplification is an assumption, not a consequence of a shared instrument.[ref]
Evaluation of measurement data — Guide to the expression of uncertainty in measurement by JCGM. Section 5.2, correlated-input propagation and sensitivity coefficients
Reweighting also changes what is measured. Subtracting from cancels when both sensitivities are nonzero, but estimates , not generally the original frequency difference. Achieving cancellation is insufficient unless the resulting estimator still answers the intended physical question.
A controlled reference perturbation can estimate the slopes and while the true spectral features are held stable. Their difference predicts leakage. Spectral offset identifiability asks whether the available observations can distinguish a real change of frequency separation from this leakage; one observed difference alone cannot determine both an unknown physical change and an unknown offset. Reference features, controlled perturbations or independently established response coefficients supply additional information.
Spectral offset identifiability asks whether the available spectral observations can distinguish a physical frequency change from an instrumental offset. Identifiability is a property of the measurement model and its observations. Reducing random noise cannot by itself separate quantities that always enter the model in the same combination.
For an original local model, let an observed change bewhere is the physical change, an unknown reference offset and a known sensitivity. If , every replacementgives the same . The single observation therefore does not identify both unknowns. With and , and in compatible frequency units are observationally indistinguishable, even without measurement noise.
Reference-assisted spectral offset separation adds an observation from a stable reference with known nonzero sensitivity . The combined linear system isIts determinant is , so the two ideal unknowns are identifiable when . The reference must genuinely exclude the physical change represented by ; otherwise the assumed second equation is wrong.
Identifiability also differs from precision. For fixed known , correlated-input uncertainty propagation givesA very weak reference response can make the identified estimate noisy. Uncertainty in the sensitivities requires additional terms. The primary GUM supplies the measurement-model and covariance framework; the rank and invariance arguments above are explicit derivations for this stated spectral model.[ref]
An experiment can test the separation by measuring stable reference features and the sample while deliberately changing the assigned axis offset. Repeated spectra characterize noise; reference perturbations characterize sensitivity. Neither exercise establishes spectral offset identifiability unless the observations distinguish the competing physical and instrumental changes.
Reference-assisted spectral offset separation uses an additional reference observation to distinguish a physical spectral change from an instrumental offset. The reference must respond to the offset in a known way while excluding, or separately modelling, the sample’s physical change. Merely collecting another signal does not guarantee spectral offset identifiability.
For a local linear measurement model, writewhere is the desired change, the shared offset and . With known sensitivities and zero-mean residual errors,removes the offset contribution. This is a subtraction of modelled responses, not a claim that the uncorrected sample and reference frequencies are physically identical.
An original example uses , and , giving . A reference may have a much larger offset response than the sample; the response ratio supplies the necessary conversion.
The uncertainty of a reference-corrected spectral change depends on joint errors. For fixed ,With , and correlation , this gives approximately ; assuming independence instead gives . The primary GUM supplies the correlated-input propagation rule used in this calculation.[ref]
If is itself estimated, its uncertainty also matters. Under an additional independent-input first-order approximation, the contribution is . Large uncertainties, correlations with the fitted ratio or nonlinear responses require a more complete analysis.
A controlled experiment records stable reference features through the same acquisition and processing chain while deliberately perturbing the instrumental offset. This can estimate and test whether the response is linear. Sample/reference timing must also match the assumed offset: a drifting offset at different acquisition times can leave an uncorrected residual. Repeated paired data, rather than unrelated error estimates, are needed to characterize the relevant covariance.
For reference-assisted spectral offset separation, the corrected estimate depends on the measured sample change , reference change and sensitivity ratio . Its measurement uncertainty must include their joint variation, including uncertainty in a fitted sensitivity ratio. Subtracting a reference can reduce shared noise while adding independent noise.
The gradient of the measurement model with respect to is . If is their covariance matrix, first-order uncertainty propagation gives , orThe signs follow from the sensitivity coefficients, rather than a rule that all correlation reduces uncertainty. This is an application of the correlated-input propagation law in the primary GUM.[ref]
Evaluation of measurement data — Guide to the expression of uncertainty in measurement by JCGM. Section 5.2, equations (13)–(15), correlated input quantities
For an original independent-input example, take , , , , and . Then andTreating as exact would give only . A large reference response amplifies uncertainty in its conversion ratio, even when that response is measured precisely.
For mutually independent random inputs, the exact product variance also includes :where on the right denote the input means. It follows by expanding . This term is omitted by linearization. In the example it changes the standard uncertainty to about .
Experimentally, use paired sample/reference spectra to estimate shared covariance, and independent sensitivity measurements to test the proposed value of . If the same spectra estimate both and the correction, independence cannot simply be assumed. Drift between acquisitions or a nonlinear reference response is a measurement model problem as well as a precision problem.
A vibrational photon echo is a coherently emitted infrared signal in which a pulse sequence allows contributions from different vibrational frequencies to rephase. It probes vibrational coherence and its loss. Rephasing of optical phase is different from physically reversing a molecular reaction or restoring a lost population.
An original ideal model makes the phase cancellation explicit. Suppose a group of oscillators has angular-frequency offsets distributed with density . A rephasing pathway accumulates phase during an initial interval and during detection. Its static-offset contribution isFor a zero-mean normal distribution of offsets with standard deviation ,At , the offset phases cancel. With stipulated and , the normalized static-offset factor is at and1 at rephasing. This ideal cancellation does not remove other sources of coherence decay or guarantee the same signal shape in a real experiment.
A primary peptide experiment used phase-controlled femtosecond pulses and phase matching to isolate vibrational echoes. Heterodyne detection measured their phase and amplitude, enabling two-dimensional infrared spectroscopy as pulse delays were varied.[ref]
The chosen pathway and pulse geometry matter: not every nonlinear infrared signal is an echo. Comparing delay-dependent amplitudes and phases can separate recoverable frequency dispersion from other dynamical effects, but interpretation needs a response model, pulse characterization and an account of the sample's changing environment.
The 2D IR waiting time is the controlled interval between excitation-related coherence evolution and later detection-related evolution in a two-dimensional infrared spectroscopy pulse sequence. It is often denoted or . Varying it asks how the sample's response changes between preparation and observation; it is distinct from the delays transformed into the two frequency coordinates.
A primary phase-controlled peptide experiment measured vibrational echo phase and amplitude as multiple pulse delays varied, providing a physical implementation of this multidelay spectroscopy.[ref]
Two-dimensional infrared spectroscopy of peptides by phase-controlled femtosecond vibrational photon echoes by M. C. Asplund and others. Abstract, multiple pulse delays and phase-sensitive detection.
The exact meaning of a fitted waiting-time change depends on which response pathways the experiment selects.An original illustrative model separates population loss from growth of a transferred component. Let total observable excited-state population decay as and let a transferred fraction grow as . Stipulate matched response factors and signalsWith and , the cross signal is approximately0.233 at but0.117 at . Meanwhile the transferred fraction increases from0.632 to0.865. A decreasing raw 2D IR cross peak need not imply a decreasing transferred fraction.
Under these particular assumptions, 2D IR population normalization gives . The raw cross signal reaches its maximum atwhich is approximately1.39ps here. This is an explicitly stipulated kinetic example, not a universal normalization rule or an inference of exchange from one peak.
At very short intervals, pulse overlap and coherent effects can matter; at long intervals, vibrational relaxation can leave little signal. Report pulse timing, normalization and the response model when assigning a waiting-time dependence to molecular dynamics.
2D IR population normalization relates selected amplitudes in two-dimensional infrared spectroscopy to a population fraction while accounting for factors shared between the signals. It is a measurement-model operation: a normalized ratio is not automatically a molecular probability.
Suppose a positive signal component obeys for a transferred population and for the remaining population. Here is a common survival factor, the fraction of interest, and the respective response factors. The raw ratio isIt equals only when the response factors match. Solving instead gives the 2D IR response-factor correctionFor an original example, , and yield . Substituting those response factors into the inverse relation recovers 0.5. Reading the uncorrected ratio as a population would overestimate it.
Common multiplicative gain cancels from the ratio, but common additive background generally does not. Stipulated signals give ; doubling both preserves 0.6, while adding 1 to each gives .
Uncertainty also depends on shared inputs. For ,This follows from correlated-input uncertainty propagation.[ref]
Evaluation of measurement data — Guide to the expression of uncertainty in measurement by JCGM. §5.2, covariance terms and sensitivity coefficients.
Small denominators and uncertain backgrounds can make the ratio unstable.Real 2D IR cross peaks can have signed or complex response contributions. Normalizing such values requires defining the fitted signal component and demonstrating its connection to populations. A waiting-time-dependent cross peak is an observation to model, rather than a population count supplied directly by the spectrometer.
2D IR response-factor correction adjusts a measured signal ratio when the spectral components being compared do not produce equal response per unit population. It is part of 2D IR population normalization, rather than a rule that every 2D IR cross peak directly counts transferred molecules.
For positive fitted components, suppose and , where is the fraction of interest and is a shared population-survival factor. Define the 2D IR response-factor ratio . The measured fraction obeysThe second expression follows by solving the first; requires in this model. An original example uses , and . The raw ratio is , while the corrected fraction is .
The effect is especially transparent on an odds scale:At , a small relative error of in changes the inferred fraction by about in magnitude to first order. Using instead of in the example gives . A response factor that is uncertain should contribute to uncertainty propagation, rather than being treated as an exact correction.[ref]
Evaluation of measurement data — Guide to the expression of uncertainty in measurement by JCGM. Sections 5.1–5.2, sensitivity coefficients and correlated inputs
A useful experimental check measures a suitable reference under the same pulse, detection and fitting conditions, and asks whether the inferred response ratio transfers to the unknown sample. Common multiplicative gain cancels; additive background, differing response pathways or a changing ratio need additional modeling. Primary phase-sensitive peptide measurements recorded both phase and amplitude of vibrational photon echoes, illustrating why the fitted response component must be specified.[ref]
Two-dimensional infrared spectroscopy of peptides by phase-controlled femtosecond vibrational photon echoes by M. C. Asplund and others. Abstract, phase matching and heterodyne measurement of echo phase and amplitude
Signed or complex spectral components do not satisfy the positive-component population model merely because they have been divided by a sum.The 2D IR response-factor ratio compares the response per unit population of two specified spectral components. In the positive-component model used for 2D IR response-factor correction,Here is the population fraction of interest and a shared survival factor. The 2D IR response-factor ratio is a property of this measurement model and its conditions, rather than a universal property of every 2D IR cross peak.
An independently characterized reference can identify if the model applies. For reference fraction ,For an original example, , and give . An unknown with and then has corrected fraction , whereas its raw 2D IR population normalization ratio is .
Using an unknown fraction to estimate and then using that same to recover the fraction would be circular. The reference needs independent information. Moreover, 2D IR response-factor transferability must be tested when pulse spectrum, polarization, 2D IR waiting time, detection or fitted response pathway changes. A common multiplicative gain cancels from ; a component-specific gain does not.
First-order uncertainty propagation is transparent on the logarithmic scale:At , a standard uncertainty in the reference fraction alone contributes about to the relative standard uncertainty of . Fractions near zero or one make that sensitivity larger. Shared inputs require covariance terms, rather than independent addition of every uncertainty contribution.[ref]
Evaluation of measurement data — Guide to the expression of uncertainty in measurement by JCGM. Sections 5.1–5.2, sensitivity coefficients and correlated inputs
A phase-resolved experiment also needs a stated choice of real, imaginary or fitted component: the primary peptide experiment recorded phases and amplitudes of vibrational photon echoes.[ref]
Two-dimensional infrared spectroscopy of peptides by phase-controlled femtosecond vibrational photon echoes by M. C. Asplund and others. Abstract, phase-controlled echoes and heterodyne readout
A ratio of signed or complex values does not automatically satisfy the positive population model.2D IR response-factor transferability means that a response ratio characterized under one set of conditions remains applicable to another measurement. It is an assumption to test when using a 2D IR response-factor ratio to infer populations from spectral amplitudes. A common detector gain can cancel from a ratio, while different gains or optical responses for its two components do not.
In the positive-component measurement model, writeThe shared survival factor cancels. If conditions change the responses to and , then . Equality of and is a substantive requirement, not a consequence of both signals coming from the same instrument.
An original error calculation supposes a reference established , but the unknown measurement actually has . For true population fraction , its observed amplitude ratio is . Using the old response ratio givesan overestimate even with noiseless detection. More generally, with ,A small response-ratio mismatch therefore gives .
The optical reason to check transfer is visible in the primary response theory: different pathways contain different transition-dipole products, and their amplitudes include relaxation factors. Pulse bandwidth, polarization and the selected 2D IR waiting time help determine which response is recorded.[ref]
Frequency-frequency correlation functions and apodization in two-dimensional infrared vibrational echo spectroscopy: A new approach by Kyungwon Kwak and others. §II.A, Eqs. 5–7 and response pathways
This does not establish a universal two-population model for every 2D IR cross peak.A 2D IR response-factor validation mixture supplies independently known composition under the new conditions. Test whether the old response ratio predicts its amplitudes, and repeat over an appropriate composition range if matrix effects or nonlinearity are plausible. Reusing an unknown's fitted fraction to validate the factor that produced it would be circular.
Retain the amplitude definition and phase convention as well as the instrument settings. The phase-controlled peptide experiment measures phases and amplitudes, so a signed or complex component need not obey the positive-component equations above.[ref]
Two-dimensional infrared spectroscopy of peptides by phase-controlled femtosecond vibrational photon echoes by M. C. Asplund and others. Abstract and experimental method, phase matching and heterodyne readout
Successful transfer concerns a specified model and observable, not every intensity visible in a spectrum.A 2D IR response-factor validation mixture has independently established information about the spectral components of interest and is measured to test a response model under specified conditions. It tests 2D IR response-factor transferability rather than merely reproducing the same fit that supplied an unknown population.
In the positive-component model , , the 2D IR response-factor ratio predictsFor , independently known fractions predict ratios . Testing several fractions probes more of the response curve than one agreement at a single composition.
An original noiseless example exposes a background error. Suppose the actual measured components areIgnoring the additive terms and inferring a response ratio separately from each mixture givesAt , the results are approximately . Their composition dependence contradicts a single uncorrected response ratio even without measurement noise. Subtracting the stipulated background from each channel recovers the common value in this constructed model.
Independent composition information must also concern the quantity used in the model. A prepared mole fraction is not automatically the excited-state population fraction if the components absorb or are excited differently. Sample preparation, spectral assignment and the response model together determine whether a mixture is an appropriate validation reference.
A useful controlled comparison retains pulse spectrum, polarization, 2D IR waiting time, phase convention and amplitude-fitting rule, then changes the condition whose transferability is being tested. Blank measurements and mixtures not used to estimate provide distinct checks. The primary response theory includes pathway-dependent transition-dipole products and relaxation factors, motivating this care with the selected observable.[ref]
Composition and background estimates carry measurement uncertainty. The uncertainty analysis should include shared inputs and covariance, rather than interpreting every departure from the curve as a new molecular mechanism.[ref]
Evaluation of measurement data — Guide to the expression of uncertainty in measurement by JCGM. Sections 5.1–5.2, sensitivity coefficients and correlated inputs
A 2D IR cross peak is an off-diagonal spectral feature correlating excitation and detection frequencies that differ in a two-dimensional infrared spectroscopy experiment. Its location is a relation between frequency coordinates, not a second physical location in the sample. Such features can reveal interactions between vibrations, but their interpretation depends on the measured response and timing.
For a concrete primary example, phase-controlled peptide spectra showed cross peaks interpreted as coupling between amide-I modes. The experiment measured the phase as well as amplitude of vibrational photon echoes, supplying information beyond an unsigned intensity map.[ref]
An original positive-envelope model illustrates 2D IR diagonal-peak overlap. Let and put two purely diagonal features at frequencies and :At the off-diagonal coordinate , this model givesFor , and , all in , the value is approximately0.0879. The diagonal value is approximately1.0019. Thus a nonzero off-diagonal pixel about8.8% of the diagonal height can arise solely from broad diagonal tails, with no separate cross feature in the stipulated model.
A real response can have positive and negative components, so this envelope is not a universal line-shape formula. It demonstrates why a peak assignment needs a fitted background/overlap model and appropriate spectral resolution, rather than the observation of any off-diagonal intensity.
Changing 2D IR waiting time, polarization or sample conditions can test proposed mechanisms. Growth or decay of a 2D IR cross peak alone does not uniquely establish chemical conversion: coupling, relaxation and changes in response factors can contribute. Interpret the feature jointly with the rest of the spectrum and the experimental pulse sequence.
2D IR diagonal-peak overlap is the contribution of extended diagonal spectral features to regions where an off-diagonal signal might be assigned in two-dimensional infrared spectroscopy. A nonzero value at an off-diagonal coordinate is not sufficient evidence for a separate 2D IR cross peak.
Peak orientation matters as well as width. An original positive-envelope model for a diagonal feature centered at isHere specifies 2D IR line-shape correlation. It is a shape parameter in this illustrative model, not automatically a measured population correlation. Let for two diagonal features, with . At the off-diagonal coordinate ,For original values , and , all in , the off-diagonal amplitude is approximately 0.0879 at , but only 0.000340 at . Both models have the same individual marginal width. An overlap estimate based only on that width can therefore miss a substantial shape dependence.
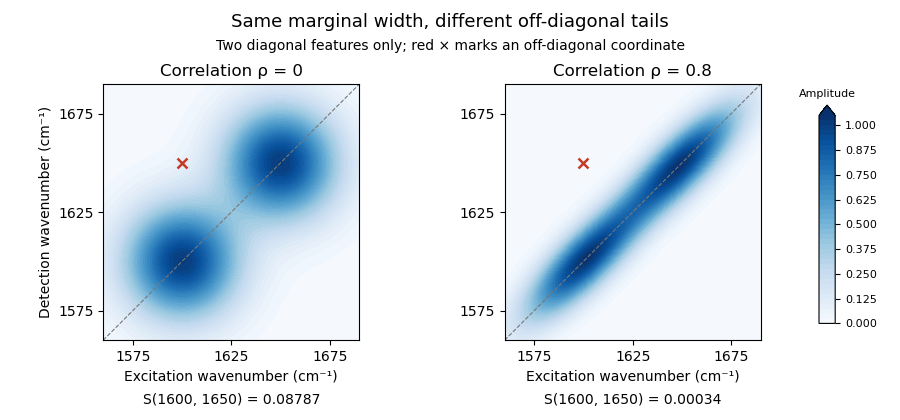
Original two-diagonal-feature model: changing 2D IR line-shape correlation changes off-diagonal tails without adding a 2D IR cross peak
. Details. The toy envelope is positive. Actual coherent spectra can contain signed features, so fitting and subtraction must respect the measured response and phase convention. The primary phase-controlled peptide experiment used phase and amplitude information to interpret mode-coupling cross peaks.[ref]
Useful tests vary the fit region, include plausible neighboring features and compare the proposed cross signal across 2D IR waiting times and other controlled conditions. A residual can indicate a missing feature, an unsuitable diagonal line shape or an unmodeled background. Its persistence alone does not uniquely select one molecular explanation.
2D IR line-shape correlation describes how excitation- and detection-frequency coordinates vary together within a spectral feature in two-dimensional infrared spectroscopy. An elongated feature can retain a relation between the initially selected frequency and the frequency read out later. Its shape can change with 2D IR waiting time, but a fitted geometric parameter needs a response model before it is assigned a molecular dynamical meaning.
An original Gaussian-envelope example makes the geometry precise. Let centered frequency offsets be , with widths and correlation coefficient :The associated covariance matrix isAt fixed , maximizing over gives the lineThus a 2D IR center-line slope equals only for this orientation with equal widths in compatible frequency coordinates. For stipulated widths and in and , the slope is . A slope greater than one is not an impossible correlation coefficient; the two quantities have different width dependence.
For equal widths , rotated coordinates and have variances and . At , the diagonal-to-antidiagonal width ratio is . This geometric identity also helps explain why 2D IR diagonal-peak overlap depends on orientation as well as marginal width.
A 2007 primary analysis by Kyungwon Kwak, Sungnam Park, Ilya J. Finkelstein and M. D. Fayer extracted a waiting-time-dependent observable from maxima of spectral slices and tested it against calculated spectra with known frequency dynamics. Its center-line convention explicitly depends on the direction of the slices.[ref]
Frequency-frequency correlation functions and apodization in two-dimensional infrared vibrational echo spectroscopy: A new approach by Kyungwon Kwak and others. Abstract, Introduction and Section II.B, center-line definition and calculated-spectrum tests
A vibrational frequency-frequency correlation function instead concerns frequency fluctuations at two times. Under an appropriate stationary model, its normalized form is with . Relating it to spectral geometry requires the pulse sequence, broadening model and slope convention. Multiple overlapping peaks, signed responses and finite acquisition windows must be addressed before interpreting a change of shape as a unique molecular time scale.
A vibrational frequency-frequency correlation function describes how deviations of a vibrational transition frequency at two times vary together. For a real stationary process with mean , defineThis is an autocovariance function. It has squared-frequency units, and is the variance of the frequency fluctuations. If , the normalized quantity starts at one. Normalizing removes the fluctuation amplitude, so two systems can have the same but different .
Vibrational spectral diffusion is the associated evolution of transition frequencies in their changing molecular surroundings. Decay of their correlation need not mean loss of vibrational population.
An original exponential example isWith and , and , while . The mean transition frequency has not been specified by these fluctuation parameters.
The function also connects a frequency model with coherence decay. For a zero-mean classical Gaussian process, neglect population loss and other broadening, and let accumulated phase be . Its Gaussian average givesFor the exponential model,Thus the chosen example has and coherence factor about . These are stipulated model values, not an experimental fit.
The 2007 primary analysis relates vibrational spectral diffusion to changing 2D IR line-shape correlation. Three controlled infrared pulses label an initial frequency and later read out the response, with the intervening 2D IR waiting time varied.[ref]
Frequency-frequency correlation functions and apodization in two-dimensional infrared vibrational echo spectroscopy: A new approach by Kyungwon Kwak and others. Introduction, pulse delays, frequency labeling and frequency-frequency correlation function
A measured 2D IR center-line slope can constrain such dynamics under appropriate response approximations; it is not a direct time trace of each molecule's frequency. Vibrational relaxation and instrumental effects need separate consideration.Vibrational spectral diffusion is the evolution of a molecular vibrational transition frequency as its surroundings or configuration fluctuate. It concerns frequency memory within a vibrational response, whereas vibrational relaxation concerns loss or redistribution of vibrational excitation. Their characteristic times need not coincide.
In two-dimensional infrared spectroscopy, an initial frequency label is followed by an independently varied 2D IR waiting time and a later frequency-sensitive readout. The vibrational frequency-frequency correlation function models the relation between those earlier and later frequencies, while a 2D IR center-line slope summarizes part of the measured spectral geometry under specified response assumptions.[ref]
Frequency-frequency correlation functions and apodization in two-dimensional infrared vibrational echo spectroscopy: A new approach by Kyungwon Kwak and others. Introduction and §II.A, frequency labeling, population interval and correlation-function model
An original two-state vibrational frequency memory model has equally populated environments with spectroscopic frequencies and . Suppose switching in either direction follows a continuous-time Markov chain with rate . With a sign variable , the probability of a flip during is ; each flip reverses its frequency deviation. Hence the normalized sign memory satisfiesThe memory time is , not the mean residence time in either state. Returning to the initial state contributes to the measured memory, so these two time scales describe different events.
For stipulated transitions at 1000 and with , the mean is , the variance is and the memory time is 2 ps. At 2 ps the correlation is . The probability of occupying the initial environment is , including trajectories with an even number of switches.
If an independent vibrational population in this illustrative model decays with lifetime 10 ps, its surviving fraction at 2 ps is , while normalized frequency memory is only 0.368. This explicitly separates population survival from frequency decorrelation. These are model values, not measured solvent or vibrational constants.
The model uses spectroscopic wavenumber, not angular frequency. Converting to an angular-frequency correlation multiplies by when the speed of light uses compatible length units. Mixing these conventions changes the fluctuation amplitude without changing the switching time.
To test the interpretation, acquire spectra at several waiting times, characterize pulse timing and phase, and assess overlap, population decay and instrument response. A decaying spectral statistic can constrain frequency fluctuations, but the same observed decay may fit more than one microscopic environment model. The two-state calculation is a reusable mechanism example, not a universal explanation of spectral diffusion.
Two-state vibrational frequency memory models a probe that switches between two frequency environments while retaining a time-dependent relation to its earlier state. It distinguishes state occupancy, a vibrational frequency-state residence time and decay of the vibrational frequency-frequency correlation function.
Let a continuous-time Markov chain switch from state A to B at rate and from B to A at rate . Its stationary probabilities areFor spectroscopic wavenumbers , the stationary mean is and the variance isThe centered state values form a decaying mode of the two-state transition generator, with rate . Consequently,The vibrational frequency-state residence times are instead in A and in B. Correlation decay includes paths that leave and return, so it is not identical to survival without a switch.
Starting in A, the probability of being in A later isAt it is about , while the probability of never leaving A is only . Their difference includes return trajectories. At long times, the stationary probability of matching the initial state approaches , while normalized frequency covariance tends to zero. A nonzero same-state baseline therefore need not mean persistent covariance.
Primary two-dimensional infrared spectroscopy theory connects waiting-time-dependent spectral geometry with frequency memory under response approximations.[ref]
Frequency-frequency correlation functions and apodization in two-dimensional infrared vibrational echo spectroscopy: A new approach by Kyungwon Kwak and others. Introduction and Section II.A, frequency labeling and correlation-function description
The rates and wavenumbers above are illustrative. A real assignment also needs spectral resolution, pulse characterization and a population-relaxation model; agreement with one exponential decay does not uniquely establish two microscopic states.A vibrational frequency-state residence time is the duration a probe remains in a defined frequency environment before leaving it. It differs from the time over which the probe's frequency retains statistical memory, and from a vibrational relaxation lifetime describing excitation loss.
For a state with constant exit rate , let be the probability of no exit by time . The small-interval exit model givesThe residence-time density is , with mean and variance . The exponential distribution is memoryless: . These conclusions require a constant-rate model, rather than following from the name of a spectral state alone.
For a two-state continuous-time Markov chain with A-to-B rate and B-to-A rate , the mean residence times are and . The normalized two-state vibrational frequency memory instead decays as . Returning after an exit contributes to occupancy and correlation but not to survival without an exit.
An original symmetric example uses . Mean residence time is , while frequency-memory time is . Starting in A, the probability of being in A after iswhereas no-exit probability is . At these are about and .
This produces vibrational residence-time sampling bias if a same-state observation is treated as proof that no exit occurred. Using as an exit-rate estimate gives about , below the stipulated . Faster time sampling can resolve more excursions, but the estimator must still account for missed transitions and state-assignment uncertainty.
In two-dimensional infrared spectroscopy, the 2D IR waiting time separates initial frequency labeling from later response readout. Primary response theory relates such measurements to frequency memory under explicit approximations.[ref]
Frequency-frequency correlation functions and apodization in two-dimensional infrared vibrational echo spectroscopy: A new approach by Kyungwon Kwak and others. Introduction, pulse timing and frequency-frequency correlation function
A fitted memory decay is therefore not automatically a directly measured single-state residence time. Nonexponential survival can require additional states, time-dependent rates or a different physical model.Vibrational residence-time sampling bias occurs when finite-time observations miss exits and returns between frequency states, or when imperfect state assignment creates apparent transitions. The resulting apparent dwell behavior need not equal the underlying vibrational frequency-state residence time.
For an original symmetric two-state continuous-time Markov chain with exit rate in either state, the probability that two samples separated by differ isIt counts an odd number of switches, not every switch. The expected observed-change count per observation time is thereforewhich approaches only as the interval becomes small. At long intervals it approaches even if the true process switches much faster.
With and , and . Treating same-state occupancy as no-exit survival gives a different naive estimate, . Both differ from the stipulated true rate.
Under this specific symmetric model, inversion givesUsing the exact model probability recovers the chosen rate. An estimated probability near makes the inverse very sensitive to uncertainty, and the nonlinear inverse is not automatically an unbiased finite-sample estimator.
Misclassification can bias in the opposite direction. If a physically unchanged two-state label is independently reversed with probability at each observation, the apparent transition probability is . At it is despite no physical transition. Faster sampling alone does not remove this labeling error.
A useful measurement comparison varies sampling interval and checks state assignment against suitable references. In two-dimensional infrared spectroscopy, a 2D IR waiting time separates frequency labeling and later response, but spectra are not automatically direct individual-state trajectories.[ref]
Frequency-frequency correlation functions and apodization in two-dimensional infrared vibrational echo spectroscopy: A new approach by Kyungwon Kwak and others. Introduction and Section II, pulse timing and response model
Relating a spectral memory statistic to residence times requires a physical observation model as well as a transition model.A 2D IR center-line slope is obtained by locating peak maxima in a sequence of slices through a two-dimensional infrared spectroscopy feature, then fitting the line traced by those maxima. Its definition includes which coordinate is held fixed, which coordinate is maximized and which coordinate is plotted vertically. Those choices constitute the 2D IR slope-axis convention.
Let denote excitation-frequency offset and detection-frequency offset. For a positive bivariate Gaussian envelope with widths and correlation coefficient , completing the square gives the slice maximumReversing the slicing direction givesThese conditional-maximum slopes are generally not reciprocals. For an original example with , and in compatible frequency units, they are and . Taking would not produce the reverse-slice value. Their product is for this particular Gaussian model.
A simple linear regression estimator from slice maxima isThis fits the maxima, rather than calculating a spectral-intensity-weighted covariance. Weak slices, overlapping features or asymmetric peaks can move their maxima and change the fitted line. The fitting interval, baseline treatment and slice direction therefore belong with the reported value.
The primary 2007 method uses a specified inverse-slope convention and tests its relation to the vibrational frequency-frequency correlation function using calculated spectra.[ref]
Frequency-frequency correlation functions and apodization in two-dimensional infrared vibrational echo spectroscopy: A new approach by Kyungwon Kwak and others. Abstract and Section II.B, center-line definition and inverse-slope convention
Its convention must be read before comparing its observable with a raw geometric slope or a software output.For a validated response model, measuring the slope at successive 2D IR waiting times can reveal changing 2D IR line-shape correlation. A single fit does not by itself identify a molecular relaxation time, and finite time windows or signed spectral contributions require attention before treating a spectral feature as a probability envelope.
A 2D IR slope-axis convention specifies which frequency coordinate is held fixed when taking spectral slices, which coordinate supplies each slice maximum, and which coordinate is plotted vertically. It also specifies whether the reported 2D IR center-line slope is that geometric slope or its inverse. These choices are needed before comparing slopes from two-dimensional infrared spectroscopy experiments or software.
Let be an excitation-frequency offset and a detection-frequency offset. An original positive bivariate normal distribution envelope with widths and correlation coefficient has conditional maximaThe first relation maximizes over at fixed ; the second maximizes over at fixed . They trace different lines unless the distribution is degenerate. Changing the slicing direction is not the same operation as taking the reciprocal of an already fitted line's slope.
For , and in matching frequency units, the first slope is 1.2. The second conditional slope is 0.3. Plotting that second set of maxima with vertical gives geometric slope ; its inverse is 0.3, not 1.2. Both sets came from the same stipulated band.
The 2007 primary method takes cuts parallel to the initial-frequency axis and defines its reported quantity through an inverse-slope convention. Its relationship with a vibrational frequency-frequency correlation function is established within the paper's response approximations.[ref]
Frequency-frequency correlation functions and apodization in two-dimensional infrared vibrational echo spectroscopy: A new approach by Kyungwon Kwak and others. Abstract and §II.B, center-line construction and inverse slope
A generic graphing routine's slope label does not identify that observable.Units can change a geometric slope without changing the underlying correlation coefficient. Under coordinate scalings and , a fitted slope becomes . Expressing only the vertical axis in GHz instead of THz multiplies its numerical slope by 1000. Using the same conversion on both axes leaves the slope unchanged. Reciprocal wavelength coordinates require their own transformation rather than a constant rescaling over an arbitrary bandwidth.
Report axis labels, units, slice direction, fit range and inversion convention with the result. At in the ideal envelope, the maxima at fixed lie on : that line is vertical even though its inverse slope tends to zero. A failed finite geometric fit near this limit need not mean molecular frequency memory increased. Noise, overlapping signed bands and finite fitting ranges require separate checks.
Articles by others on the same topic


Two-dimensional infrared (2D IR) spectroscopy is a powerful analytical technique used to investigate the dynamics and interactions of molecular systems. It combines the principles of traditional infrared spectroscopy with two-dimensional data analysis, allowing for a more detailed characterization of molecular vibrations, interactions, and conformations.