Assume well-mixed populations, exponential prey growth without density regulation, predator mortality without food, and bilinear encounters. The Lotka-Volterra predator-prey model isIts coexistence equilibrium is , . The Jacobian matrix there has eigenvalues , so it is a center equilibrium rather than an attracting equilibrium. More globally,is a first integral: substituting the two equations into gives zero. It has a strict minimum at coexistence, diverges at the boundary and at infinity, and its other positive-quadrant level sets are closed periodic orbits. Small oscillations have frequency . In scaled variables and , the linearization is , .
Model an annual fixed quota as a pulse removing prey, , , with . Between pulses the original equations apply. In logarithmic coordinates the flow has zero divergence, but the pulse has determinant . Therefore a periodic annually harvested trajectory has return map determinant greater than one, and cannot have both multipliers inside the unit circle. It is unstable, even when a leading rotation-plus-translation approximation looks neutral. Near a nonresonant small-quota forced orbit this expansion destabilizes the original center equilibrium; resonant pulse timing can add secular effects. Quotas larger than the remaining prey are not biologically realizable and require stopping the model. This is fixed-quota harvesting of Lotka-Volterra populations.
The common continuous approximation to an annual quota gives . Its positive equilibrium is , , requiring . Its Jacobian matrix has trace and determinant , so this approximation also destabilizes coexistence. At small it is an unstable focus. A quota is a larger per-capita mortality when prey are scarce, which is the adverse feedback responsible for the result.
For abundance-triggered pulses, retain the same fixed prey removal and specify the crossing direction. The exact event-triggered harvesting by conserved energy calculation isIf prey abundance triggers the event at , this increment is the same at every event. A sufficiently high prey threshold with a modest quota gives : the orbit's oscillation energy decreases until it no longer crosses the threshold, after which harvesting stops. If , then and harvesting increases oscillations. At the balanced threshold satisfying , energy is unchanged and the ideal orbit is neutral. Thus the useful rule is harvesting at high prey abundance, not simply using any fixed threshold; the equality case supplies no asymptotic restoration.
A predator threshold does not hold prey abundance fixed. On a rising-predator crossing, , so a sufficiently small harvest initially lowers ; on a falling-predator crossing , it raises . Nevertheless the rising-predator rule is not a robust stabilizer in this conservative model. Its balanced return orbit has preharvest prey abundanceobtained by equating before and after the pulse. On the return section the derivative isIts magnitude exceeds one because for . The balanced predator-triggered cycle is consequently unstable. At leading small-oscillation order its multiplier is , explaining a marginal alternation rather than stable damping. If the trigger is crossed in both directions, or a variable rather than fixed quota is used, a different return rule results and must be analyzed separately.
For conservation, the ideal model shows why fixed removals can be dangerous and why prey-based high-abundance feedback can be safer than a calendar quota or an unspecified predator threshold. These conclusions depend on the chosen pulse rule and on the absence of density dependence, refuges and stochasticity. They do not justify a universal numerical safe quota. A harvest policy needs a stopping threshold and the correct phase-dependent feedback, not merely a small nominal annual catch.
Use a basal resource , herbivore and predator , with positive encounter, conversion and mortality parameters:Only the resource has intrinsic positive growth; the consumers die without food. There is no resource carrying capacity, immigration or feeding on nonadjacent levels. These assumptions define the simplest food chain with Lotka-Volterra equations.
A positive equilibrium would require both and . Hence three-level coexistence is possible only on the special balance . The exact identityfollows by combining and . It gives a useful classification rather than incorrectly postulating a generic coexistence fixed point.
If , the top predator cannot invade the underlying resource-herbivore cycle: its average per-capita growth there is . This extinction statement can be strengthened. The boundary equilibrium has , , . With weights , , , defineDirect cancellation of paired trophic terms gives . Its level sets bound the first two species above and away from zero and bound . Integration and uniform continuity imply ; the limiting dynamics are the usual resource-herbivore closed orbits, not generally an attracting equilibrium.
If , there is a line of positive equilibria: , . Choosing any positive point on that line gives a conserved coercive version of with all three relative-entropy terms. Together with the product integral, it confines generic trajectories to closed curves, with neutral oscillations around the equilibrium line. All three can persist, but only under this finely balanced parameter condition.
If , the product identity grows without bound, so at least one of must become unbounded in the unregulated model. There is no bounded persistent three-species attractor. Finite-time blow-up is excluded by a weighted total-abundance bound whose encounter terms cancel and whose derivative is at most a constant times that abundance. The unbounded long-term behavior signals the missing biological resource regulation. The simplest odd chain does not generically permit bounded full coexistence; adding resource self-limitation changes this conclusion.
Add a fourth level usingwith the first two equations unchanged. The positive equilibrium must beThe first three entries are positive automatically. The fourth is positive exactly when . Thus the printed coexistence assertion requires adequate herbivore production to support the third and fourth levels; it is not true for every positive parameter set. The top predator supplies the extra mortality that was absent in the unbounded three-level case.
For a chain of levels, the same equations give in the interior, with the basal and top equations fixing alternating levels. The interaction matrix has zero diagonal and opposite-sign nearest-neighbor entries. Its determinant is nonzero for even and zero for odd : expansion at the end gives , with , . For even this gives a unique equilibrium candidate, but its entries still need to be positive. For odd a compatibility condition is necessary, just as for the three-level example. This is Lotka-Volterra food-chain parity. The parity result is specific to an exponential unregulated resource and nearest-neighbor trophic interactions.
Assume the equilibrium in part (a) is strictly positive and start with all four species positive. Choose weights recursively by and defineThe weighted interaction matrix is skew-symmetric matrix: for every adjacent pair . On differentiating , the equilibrium removes the constant growth terms, while paired products cancel. Thus . Each summand is nonnegative and diverges as its positive argument approaches zero or infinity. A fixed level of therefore bounds every abundance above and away from zero. This proves genuine persistence of all four species, although their dynamics need not converge to a fixed point.
Integrate each per-capita equation over and divide by . Since is bounded, the left side tends to zero. Writing , the limiting balances givewith the explicit abundances listed in part (a). The limit exists because the invertible four-level interaction matrix solves the averaged linear equations, whose residuals tend to zero. Periodicity or an ergodic assumption is not needed.
The same proof works for any even nearest-neighbor chain with a feasible positive equilibrium. It does not make all even-chain parameters feasible. Odd chains generically lack the required positive balance, and in their exceptional balanced cases the averaged equations do not uniquely fix every abundance. Resource density dependence and more general feeding interactions alter both the conservation law and the simple parity classification. Coexistence and the stated mean abundances hold under positivity of the equilibrium, not as an unconditional rule for every food chain.
Not in general. A food web with Lotka-Volterra equations has per-capita balances . If all species remain bounded above and away from zero, averaging requires with positive means. Many webs have no feasible positive solution. Even a feasible solution does not ensure stability or persistence when the weighted skew-symmetric matrix structure and its coercive first integral are absent.
A simple resource shared by two consumers already illustrates competitive exclusion principle. Suppose their per-capita equations are and . ThenIf , the ratio has exponential drift, incompatible with bounded persistence of both consumers. The lower break-even-resource consumer has the advantage; equal break-even requirements are a special neutral balance, not generic coexistence. More complicated feeding links can provide additional niches, but positive coefficients and a larger web do not by themselves guarantee that every species survives. One must test feasibility, invasion into the relevant boundary states and stability, with explicit assumptions about resource renewal and self-limitation.
Use fractions of all farms, so and . Assume farm size is fixed, contacts mix randomly, infectious periods have size-independent exponential mean , and the farm-size distribution has finite moments and . Write . The farm-size stratified SIR model with susceptibility proportional to size isHere includes the common contact-rate constant; using frequency-dependent mixing avoids introducing an extra total-farm factor. An infectious farm of any size generates infections in initially susceptible size- farms. The next-generation matrix has rank one, giving the basic reproduction numberThe comparison SIR model replaces every size by , so it has the same basic reproduction number and the same initial exponential growth rate .
As the epidemic develops, put . For negligible initial infection, . Large farms leave the susceptible class faster. If with , differentiation givesThe mean size of newly infected farms is , larger than the susceptible mean unless all sizes agree. If , thenRecovery cancels from this equation because its rate is size-independent. Starting with representative infectives, initially rises toward the size-biased mean ; later depletion of large susceptible farms lowers and usually lowers . It is incorrect to assert that the mean size of infectives must decrease from every possible initial condition. In the growing linearized mode, , so its mean is already .
The aggregate growth equation is . The falling susceptible mean adds a depletion effect absent from the homogeneous SIR model. The final size relation for an epidemic, with vanishing initial seed and attack fraction , isBy Jensen's inequality, the right side is at most , so is no larger than the homogeneous attack fraction. For a common small representative seed the illustrated epidemic is flatter and infects fewer farms; exact pointwise ordering of infected fractions for every time and arbitrary initial seed is not a consequence of equality of .
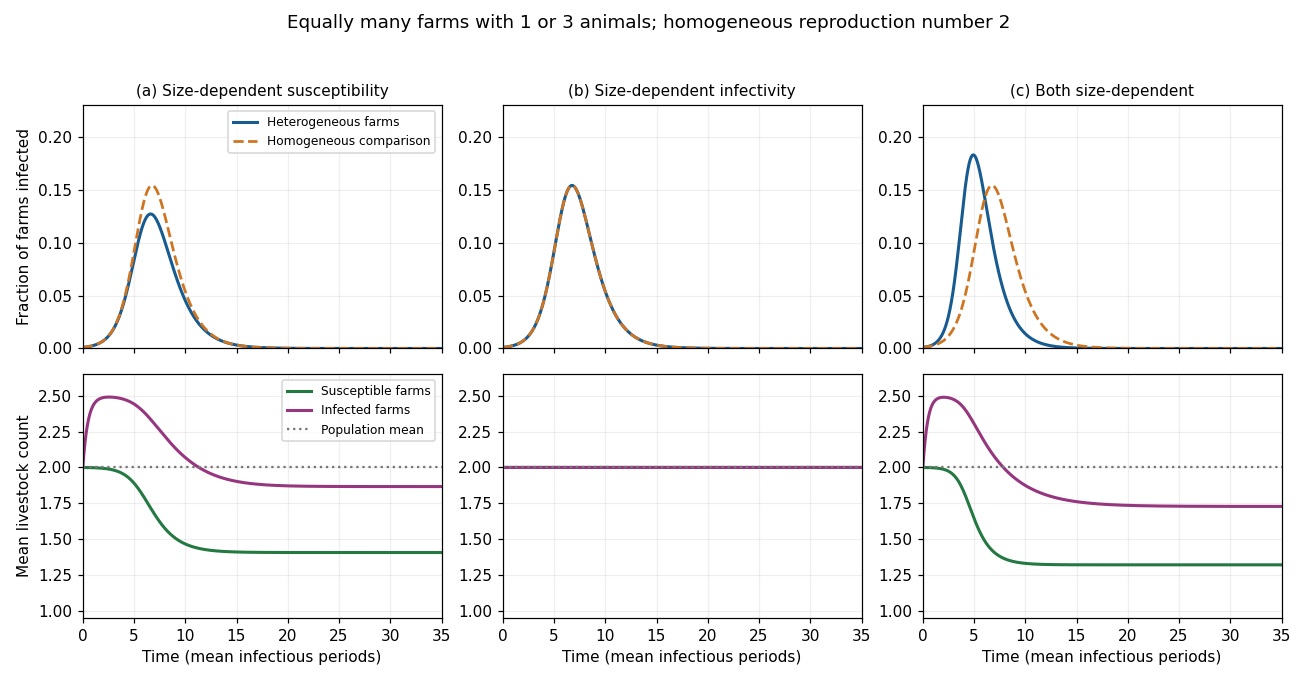
Farm-size heterogeneity: epidemic prevalence and mean livestock counts for the three transmission assumptions
. Details. 
The original sketch uses equally many farms of sizes 1 and 3, recovery rate , and a representative infected seed of . Each homogeneous comparison has . The left, middle and right columns correspond to parts (a), (b) and (c). The top row shows infected farm fractions, and the bottom row distinguishes the mean sizes of susceptible and infected farms. These are illustrative trajectories under the stated model, not universal empirical outbreak curves.
With susceptibility independent of size and infectivity proportional to size, define . The farm-size stratified SIR model isAn infectious size- farm has next-generation column . Its sole nonzero eigenvalue gives the basic reproduction numberagain equal to the comparison SIR model.
Since every susceptible farm experiences the same force of infection, all ratios remain fixed. For a representative initial seed, and , uniqueness of the differential equations gives , , for all time. Consequently andFor this representative seed, the heterogeneous and homogeneous epidemics coincide exactly, and both susceptible and infected farm means stay at . This is the overlapping middle-column sketch above, not merely an approximation based on equal reproduction numbers.
With a nonrepresentative seed the susceptible mean remains its initial value, but the infected mean need not. The influx of new infectives has the constant susceptible mean, soThus it is pulled toward while incidence continues. Infectivity variation alone does not selectively remove large susceptible farms. The exact homogeneous reduction depends on the representative initial condition; specifying only and does not determine a trajectory for every possible seed.
Both weights now equal farm size, giving the farm-size stratified SIR modelThe next-generation matrix is , whose nonzero eigenvalue isThe variance matters because large farms are both more likely to acquire infection and more effective sources afterward. This positive correlation increases the basic reproduction number relative to the homogeneous SIR model. In particular the heterogeneous population can permit invasion when the homogeneous comparison is subcritical.
Put . Apart from the negligible initial seed, . The susceptible mean obeys . For infectives, the incidence mean is again , andThe growing mode has mean , not . As the larger susceptible farms are depleted, the mean size of incoming infectives falls; the current infected mean follows it with a delay. These equations give the direction of change for any initial condition without imposing false monotonicity of the infected mean.
The weighted infected total satisfies , so the effective reproduction number declines from as the susceptible size distribution changes. The final size relation for an epidemic becomesThe right-column sketch illustrates the larger initial growth rate and preferential depletion of large farms. The homogeneous comparison has size in both transmission factors; it must not be assigned the heterogeneous second moment. The general conclusions are enhanced invasion and selective removal of large susceptible farms. The final unweighted fraction of farms infected has no universal ordering relative to the homogeneous comparison: higher transmission and stronger heterogeneity-induced depletion compete, and the displayed final-size equations determine the answer for a specified distribution. For example, with , the infinitesimal-seed attack fractions are approximately versus when the homogeneous reproduction number is , but versus when it is ; the two comparisons have opposite ordering.
The basic reproduction number is the expected value of secondary infections caused by one typical infectious individual introduced into an otherwise susceptible population. For a heterogeneous model it is the dominant eigenvalue of the next-generation operator, so the appropriate infected-type distribution is part of “typical.” A small introduction grows when and decays when , under the usual linear invasion assumptions.
The asymptotic susceptible level depends on the model. In a homogeneous SIS model with transmission rate and recovery rate , the positive endemic equilibrium has when . In a closed SIR model, however, the final size relation for an epidemic isFor an infinitesimal seed in a wholly susceptible population this becomes . The epidemic peak occurs at , and a nontrivial outbreak leaves . Thus is an endemic or instantaneous invasion threshold, not a universal final susceptible fraction.
For cooperative infections, let be the fractions carrying neither infection, only , only , and both, with sum one. Let , , assume each infection independently recovers at rate , and multiply susceptibility to the second infection by . One suitable pair of SIS models isRecovery of one infection from leads to the other single-infection class, not directly to . The probability simplex is invariant. At the disease-free equilibrium the eigenvalues are , and . Therefore failure of both isolated and low-density invasion requires , . Enhancement of secondary susceptibility does not alter these linear thresholds because coinfection is second order at introduction.
For completeness, a joint positive equilibrium must haveandTo obtain these, divide the equilibrium equations for total and total prevalence by their positive prevalences, yielding and . Substitution into the balance gives the quadratic. Its discriminant must be nonnegative and the resulting positive. These conditions test equilibrium feasibility for unequal rates; stability must also be checked rather than inferred from the mere presence of a root.
A fully explicit Allee effect results in the symmetric permitted case , . The joint equilibria haveTwo feasible positive equilibria coexist with the stable disease-free state exactly in the strict rangeIndeed at the fold the two roots coalesce, and at the smaller-prevalence branch reaches the disease-free boundary. For , roots at do not give positive singly infected fractions. Strict inequalities exclude these marginal endpoints.
Here is the stability check underlying the cooperative SIS coinfection threshold. On the symmetric subspace put and . ThenAt a positive equilibrium . Its two-dimensional Jacobian matrix hasThe higher-prevalence branch has positive determinant and negative trace, while the lower-prevalence branch is a saddle equilibrium. The remaining antisymmetric mode, from , has eigenvalue . Thus the smaller susceptible root is a stable joint endemic equilibrium, separated from the stable zero state by a finite seeding threshold. Neither infection can persist alone in this range. Strong cooperation can sustain both infections after a sufficiently large joint introduction even though neither invades from rarity. The numerical condition belongs to the specified secondary-susceptibility model; alternative cooperation mechanisms need their own threshold calculation.
Let be fractions of patches occupied by the two competitors and empty, with . Assume well-mixed propagule dispersal, equal independent fire extinction rate , colonization constants , and net replacement advantage for the poorer colonizer. The Levins metapopulation model with replacement isReplacement fluxes and cancel in the total occupancy equation. At every face the outward flux is zero or points into the patch probability simplex, so these equations preserve valid fractions. The constants include both propagule arrival and establishment probability; the model does not require unequal fire susceptibility.
Put . Solving the two occupied-patch balances and their sum constraint givesThus strict coexistence is equivalent toToo little disturbance lets the better competitor exclude the colonizer; too much disturbance favors colonization or destroys occupancy. A positive equilibrium automatically has . If the poorer colonizer cannot survive alone, coexistence can nevertheless be possible because it repeatedly replaces established patches of the better colonizer.
These bounds also have an invasion interpretation. The -only equilibrium is when ; the growth rate of rare there is , positive at the upper-bound condition above. At a feasible -only equilibrium the rare- growth is , giving the lower bound. At coexistence the occupied-patch Jacobian matrix has trace and determinant , proving local asymptotic stability. The Bendixson-Dulac criterion with multiplier gives divergence in the interior, excluding periodic competition cycles.
To specify selection, a monotone tradeoff alone is insufficient: one must say how two different trait values determine replacement. A natural model uses a trait with increasing competitive ability and decreasing colonization , and net replacement by a mutant against a resident . The resident metapopulation occupies , requiring . The rare-mutant per-capita growth, or invasion fitness, isIt is zero at . Differentiating with respect to the mutant trait gives the selective pressureHigher colonization is valuable in empty habitat; higher competitive ability is valuable where the resident already occupies patches. This is colonization-competition tradeoff invasion fitness.
At an interior candidate for a local evolutionarily stable strategy, . A strict, nondegenerate local evolutionarily stable strategy follows ifsince nearby distinct mutants then have negative invasion fitness. For a strict global evolutionarily stable strategy, every feasible distinct mutant must satisfyThis is the supporting-line condition on the tradeoff curve. At a trait boundary replace the vanishing-gradient condition by the appropriate one-sided inequality. Equality against another mutant, or zero local curvature, needs the second condition of evolutionary stability or higher-order terms; the strict tests above are sufficient and are the usual nondegenerate conditions, not a claim that every tie is unstable.
For evolutionary approach to a singular point, the selection gradient must also point toward it, locally . This is logically distinct from testing mutant fitness. In this particular additive-ability model with , negative mutant curvature implies negative becauseIf replacement instead has a general kernel , replace in by that kernel and differentiate it; monotonicity does not justify importing the displayed additive formula without that modeling assumption. The tradeoff shape, disturbance rate and replacement law together determine selection and local or global uninvadability.
Take “cousins” to mean first cousins with unrelated, outbred founders and unrelated spouses in the intervening generation. Interpret a homozygous chromosome as two copies identical by descent, and first use an idealization of independent, nonrecombining chromosome blocks. Ordinary sequence homozygosity is a different quantity: unrelated copies can have the same allele, so its probability also requires allele frequencies. Similarly, identity of an entire recombining chromosome requires a genetic map, not just a pedigree.
At a single autosomal locus, the probability of autozygosity in their child is the parents' kinship coefficient. Each of the two common grandparents contributes a path of length two from each parent, with probability . Thus the first-cousin inbreeding coefficient isIf there are independent autosomal blocks, the probability that none is autozygous is . Hence the usual 23-block idealization givesThe exponent describes the stated independent-block approximation; it is not a claim that all 23 human chromosome pairs inherit like autosomes.
Humans have 22 autosomal pairs. A son has one X and one Y and cannot have two homologous X copies. A daughter can have an autozygous X, with probability determined by the sex-linked cousin kinship calculation in part (b). Thus a sex-aware version iswhere is the probability of a daughter, or is one or zero for a specified daughter's or son's sex. The independent assortment approximation applies between the autosomes and the X. Opposite-sex parentage alone does not determine : the parents may be cousins through their mothers, their fathers or one of each. Without these pedigree and chromosome-block assumptions, the wording does not specify a unique probability of an entire homozygous human chromosome.
Use the conventional additive pedigree relatedness , where is the kinship coefficient, the probability that randomly selected gene copies from and are identical by descent. For noninbred individuals this convention gives ordinary full siblings relatedness , as required. It is important that this is not the probability of sharing at least one copy: that probability is for ordinary full siblings.
Let the parents be . They are otherwise outbred first cousins, so and . For two children , independent Mendelian segregation givesTherefore the autosomal additive relatedness isThe additional beyond comes from maternal-paternal cross-origin matches. Each child has inbreeding coefficient . The additive coefficient is a sum of descent probabilities rather than a literal bounded probability in all inbred pedigrees; a relationship normalized to self-relatedness would be . Likewise, identity-by-descent sharing of inbred full siblings gives a different matched-copy statistic. The exam's calibration to supports the additive convention, but the quoted probability wording needs this qualification.
For sex chromosomes the route of cousinship matters, not merely the fact that the parents have opposite sex. Let be the mother-cousin's parent in the shared sibling pair, and the father-cousin's parent in that pair. Let be the probability that the mother's randomly transmitted X matches the father's single X by descent. With no founder inbreeding, the possibilities areIn the first two rows the father-cousin is related through his father, whose X he does not inherit. In the third row his mother is a sister of the mother-cousin's father: the relevant brother and sister match their maternal X copies with probability , and the mother-cousin transmits her paternal X with probability , giving . In the last row the two sisters' random X gametes match with probability : their common father's X contributes , and their common mother's two X copies contribute . Transmission by the mother-cousin halves this to . This derives the sex-linked cousin kinship values.
For an unweighted comparison by chromosome pair, two daughters have sex-pair additive relatedness : their paternal X always matches, maternal X matches with probability , and the two cross-origin terms each contribute . Two sons have mean sex-pair relatedness , from an X match of probability and a common paternal Y with probability one. A daughter-son pair has sex-pair value , with the Y contribution zero. These calculations treat chromosomes as unrecombined blocks and weight X and Y equally; chromosome-length-weighted or locus-specific coefficients are different.
For two children with independent equally likely sexes, these three cases have weights . Their mean sex-pair relatedness is , so averaging the 22 autosomal pairs and the sex pair givesIf the cousins' route has , this is , slightly below . If, as an additional sampling assumption, the four parent-sex routes in the table are equally likely, and the answer is . These are different legitimate numerical corrections under different specified pedigrees. There is no unique sex-chromosome correction from opposite-sex cousinhood alone. The table and formula give it once the missing pedigree and offspring-sex information is supplied.
Assume a closed population with even size , unrelated founders, monogamous mating, nonoverlapping generations and exactly two offspring per couple. Sex labels are exchangeable for the autosomal calculation, with enough individuals of each sex to form the stipulated pairs. A randomly chosen distinct offspring pair are siblings with probability : among the pairs, exactly are within-family pairs. This fixed-family-size fact, rather than independent sampling of all parents, determines the recurrence.
First give the elementary chromosome-sharing idealization consistent with taking self-relatedness as one and ordinary sibling relatedness as . In this model each individual carries one ancestral label, passed through one randomly chosen parent. Let be mean sharing between distinct individuals, and let be the mean sharing between mates. Siblings share their parental label with probability . The average relatedness between parents from different couples isTo see the latter count, the ordered pairs from different couples are all ordered distinct pairs minus the ordered mating pairs. Each pair of parents from different couples contributes equally to the four possible cross-family offspring relationships. Combining sibling and nonsibling offspring givesWith random mating, in expectation, so givesThe mean sharing tends to one, on a generation scale of order . It is not linear indefinitely: repeated ancestry saturates the probability. This is an exact result for the stated one-label approximation, not an exact diploid formula after substantial inbreeding has developed.
Now prohibit full-sibling matings, with and representative sampling of allowed nonsibling mating pairs. The following closed recurrences assume that opposite-sex mating pairs have the same mean autosomal sharing as all nonsibling pairs; a sex-specific or assortative scheme requires separate mating means. In the exchangeable one-label model write for mean relatedness of a nonsibling pair, so . The two required recurrences areThe second relation includes sharing through grandparents and more distant ancestors; setting forever would be wrong. For , , the linear recurrence matrix isIts eigenvalues areboth of magnitude less than one for . Thus even when siblings never mate. Explicitly, , where . Sibling avoidance prevents immediate close-relative matings, but it does not make the remaining population unrelated. In a fixed-two-offspring model it need not reduce average pairwise sharing at every generation: a smaller mating mean actually increases the displayed increment in .
For a fully diploid formulation, self-kinship also changes with inbreeding and cannot remain artificially fixed. Let be mean kinship coefficient between distinct individuals, mean inbreeding coefficient, and mean kinship of mates. A sibling pair has kinship , because parental self-kinship is . Repeating the pair count yields finite-population chromosomal coancestry:Initially . For random mating take . An explicit solution is the affine matrix recurrenceIts characteristic polynomial is ; its two roots have magnitude less than one. Hence . The normalized relationship tends to one, while unnormalized additive relationship tends to two, demonstrating why that coefficient cannot always be read literally as a probability.
With sibling avoidance replace by the nonsibling mean and addThese equations compute every generation's kinship and inbreeding, including delayed consanguinity through cousins. Under random allowed mating in a connected closed population, backward ancestral gene-copy lineages have a positive chance to meet in each sufficiently long block of generations; repeated blocks force eventual coalescence. Thus sibling avoidance alone does not prevent long-term identity by descent and genetic drift. Maintaining distinct isolated breeding groups, introducing immigrants, or changing offspring-number variance gives a different model. The elementary exponential formula answers the fixed-self-sharing approximation; the diploid recurrences give the consistent inbreeding-aware answer.
Articles by others on the same topic
There are currently no matching articles.