A ribozyme is an RNA molecule whose folded structure provides catalysis of a chemical reaction. A protein enzyme builds its active site from an amino acid chain; a ribozyme builds a working active site from RNA, sometimes with metal ions and associated proteins. Base pairing can help recognize a substrate, but recognition alone does not establish catalysis: the RNA must promote chemical conversion. A passive messenger RNA template being read by a ribosome is not thereby a ribozyme. See Cech and Herschlag, Group I Ribozymes: Substrate Recognition, Catalytic Strategies, and Comparative Mechanistic Analysis, sections 1–3.
The surprise is that an information-bearing polymer can also perform chemistry. Examples include the self-splicing group I intron of Tetrahymena thermophila and the catalytic RNA component of bacterial RNase P, discussed below. The ribosome supplies an everyday example: its peptide bond-forming active site is made of ribosomal RNA, although the complete machine also contains proteins. Nissen and colleagues examined structures of a large ribosome subunit with substrate analogues and found RNA contacting those analogues at the catalytic centre. This supports an RNA active site, not a claim that isolated ribosomal RNA can carry out all of biological translation. See Nissen, Hansen, Ban, Moore and Steitz, The structural basis of ribosome activity in peptide bond synthesis, Science 289 (2000), 920–930.
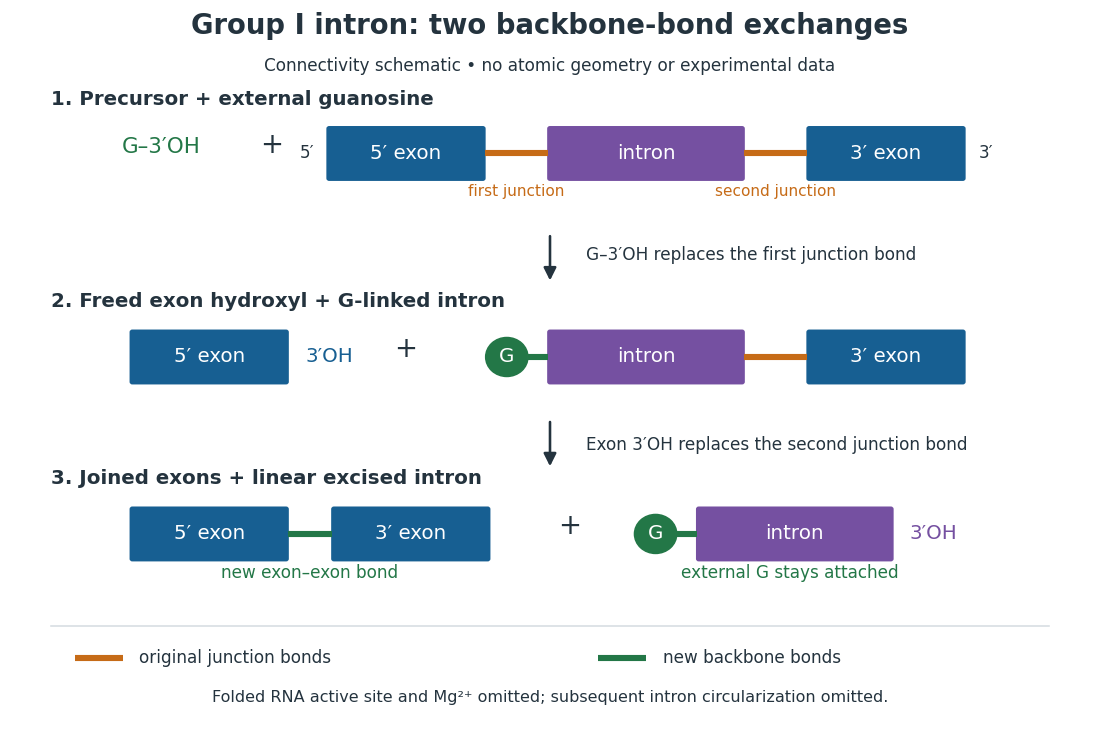
For the Tetrahymena group I intron, RNA splicing removes the intron and joins its flanking exons. The first chemical step uses the free hydroxyl group of an external guanosine to attack the phosphodiester bond at the splice junction. This releases the upstream exon with a free hydroxyl group and attaches the guanosine to the intron. That newly freed hydroxyl group then attacks the phosphodiester bond at the other splice junction, joining the exons and releasing the linear intron. Each step exchanges one backbone connection for another; it is not simply separation of base pairs. See Cech, Zaug and Grabowski, 1981: the phosphoester-transfer mechanism.
The motivating experiment began with an unexpected control. As Cech recounts, Zaug purified a ribosomal RNA precursor from Tetrahymena using a detergent and phenol extraction, then compared test tubes with and without added cell nucleus extract. Incubation in salts and nucleotides released the same amount of small intron RNA in both cases, detected by gel electrophoresis. Repetition made a missing ingredient or accidental omission less plausible. Activity also survived treatments intended to destroy proteins. Nevertheless, absence of added extract did not by itself exclude a tightly attached, resistant protein, or prove that the small RNA had been newly excised rather than released from an already processed complex. See Thomas R. Cech, 1989 Nobel lecture, pp. 655–659: the unexpected control and alternative explanations.
An independent chemical readout addressed the latter problem. Cech mixed precursor RNA with phosphorus-32-labelled guanosine bearing three phosphate groups (GTP), and used gel electrophoresis followed by exposure to X-ray film to locate the label. Label appeared in the excised intron with magnesium present; it was absent when magnesium was omitted or labelled ATP replaced the labelled guanosine reagent. This experiment is described in Cech, 1989 Nobel lecture, pp. 657–658: labelled-guanosine controls. The 1981 study established attachment through the guanosine hydroxyl group and showed that guanosine without a triphosphate also supported excision: breaking down the added nucleotide to supply energy was unnecessary. See Cech, Zaug and Grabowski, In vitro splicing of the ribosomal RNA precursor of Tetrahymena, Cell 27 (1981), 487–496: primary abstract. Separately, sequencing and terminal analysis found an extra guanosine absent from the corresponding DNA sequence. See Zaug and Cech, The intervening sequence excised from the ribosomal RNA precursor of Tetrahymena contains a 5-terminal guanosine residue not encoded by the DNA, Nucleic Acids Research 10 (1982), 2823–2838. Together, incorporation and sequence evidence support a new covalent bond, beyond a change in how RNA associates.
Kruger, Grabowski, Zaug, Sands, Gottschling and Cech's 1982 test removed dependence on a precursor purified from Tetrahymena. They cloned the intron and adjacent exons into a DNA template, made precursor RNA with purified bacterial RNA polymerase under conditions suppressing RNA splicing, and removed proteins by detergent–phenol extraction. Denaturing gel electrophoresis separated precursor species. Purified precursor was then incubated with the guanosine reagent, magnesium and salt; matched samples without this incubation supplied a precursor baseline. Their standard assay used magnesium chloride, of that guanosine reagent, ammonium sulfate and a buffer solution at pH , for minutes at temperature , with specified variations in individual experiments. Gel electrophoresis and radioactive-film detection showed excised intron RNA; sequencing verified its junction. See Kruger et al., Self-Splicing RNA, Cell 31 (1982), 147–157, figures 3–4 and Experimental Procedures.
Excision alone could leave disconnected exons. To test joining, the authors paired labelled RNA with DNA spanning the joined-exon sequence, removed exposed RNA with enzymes, and separated the protected pieces by denaturing gel electrophoresis. Spliced samples yielded protected RNA long enough to span the junction. See Kruger et al., 1982, figure 6: independent evidence for exon ligation. The experimental logic matters: an intron-sized band identifies a candidate product, while sequence and junction assays test its chemical identity.
These controls support sufficiency of the prepared RNA under the assay conditions. They do not show that every RNA self-splices, or that proteins are dispensable inside a biological cell. Many group I introns use proteins to help attain an active fold. See Cech and Herschlag, Group I Ribozymes, section 6: protein facilitation. Nor does a failed assay necessarily identify an inactive nucleotide sequence: unsuitable ions, pH, or temperature can prevent a working active site from forming.
The distinction between self-processing and repeated use became especially clear with bacterial RNase P. In 1983 Guerrier-Takada, Gardiner, Marsh, Pace and Altman separated the RNA and protein components of RNase P from two bacteria. Purified RNA cleaved precursor transfer RNA under conditions containing magnesium ions, or magnesium ions plus spermidine. The protein components alone had no catalytic activity. At – magnesium without spermidine, both components were needed. Thus the comparison demonstrated an RNA catalyst while also explaining why protein assistance could matter under other conditions. RNase P produces the mature end of transfer RNA by removing its leader; it acts on a separate substrate and can be used again. See Guerrier-Takada et al., The RNA moiety of ribonuclease P is the catalytic subunit of the enzyme, Cell 35 (1983), 849–857: primary abstract.
Self-splicing demonstrates RNA chemistry, but one precursor completing one rearrangement is not yet evidence for catalytic turnover: repeated conversion of separate substrates by a regenerated catalyst. The self-splicing precursor changes its own connectivity and incorporates external guanosine. In 1986 Zaug and Cech showed that a shortened Tetrahymena intron RNA could cleave and rejoin separate short RNA substrates in a sequence-dependent reaction. This demonstrated an RNA enzyme acting beyond its own self-processing. See Zaug and Cech, The intervening sequence RNA of Tetrahymena is an enzyme, Science 231 (1986), 470–475: primary abstract.
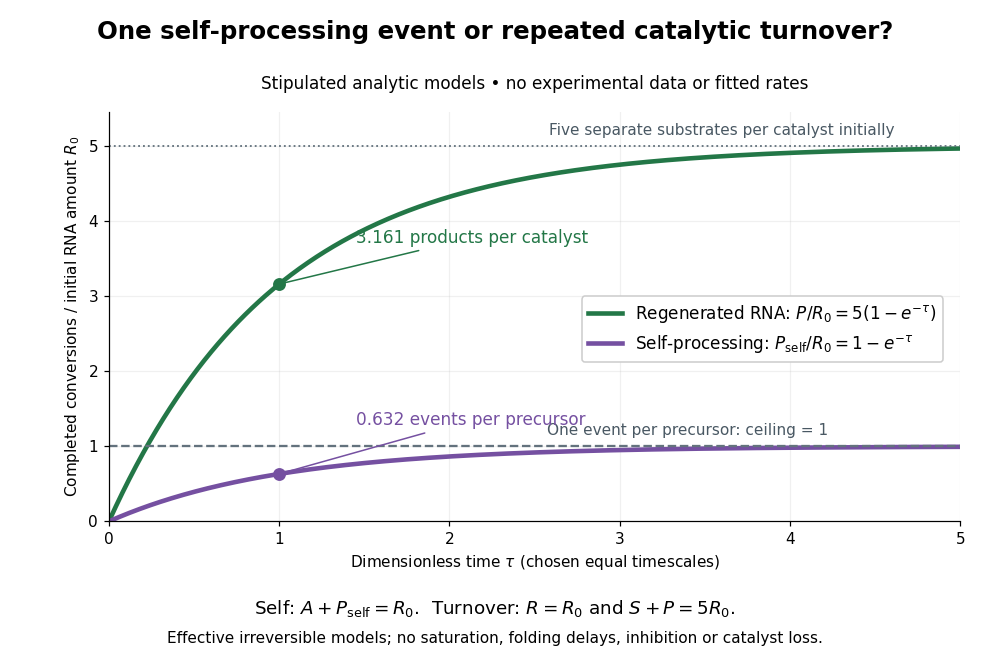
Read the second figure as a counting argument. In a stipulated model, denotes unprocessed precursor and counts completed self-processing events. Set , and . The exponential function solution gives , never exceeding one. Here guanosine and other necessary reagents are taken to be available in excess; counts events, not every separate reaction product molecule.
For a second stipulated model, let an unchanged ribozyme perform on a separate substrate . Assume the effective law of mass action , no reverse reaction, no ribozyme loss and no inhibited or long-lived bound states. Set , , and . Then and , with . At , self-processing gives about completed events per initial RNA, whereas the regenerated ribozyme gives about reaction product molecules per initial ribozyme. More than one substrate conversion per initial ribozyme, with appropriate background and identity controls, is evidence for turnover. Product accumulation below that threshold alone cannot distinguish a reusable catalyst from one-shot chemistry. These models illustrate that distinction; actual enzyme kinetics can include folding delays, binding, saturation and product release.
Articles by others on the same topic



A ribozyme is a type of RNA molecule that has the ability to act as an enzyme, catalyzing specific biochemical reactions. Unlike typical enzymes, which are usually proteins, ribozymes demonstrate that RNA can have both genetic information and catalytic activity. This property supports theories about the origin of life, particularly the RNA world hypothesis, which suggests that early life forms may have relied solely on RNA for both genetic material and enzymatic activity before the evolution of DNA and proteins.