An enzyme is a biological catalyst: it accelerates a chemical reaction and is regenerated after a successful catalytic cycle. Most enzymes are proteins, but catalytic RNA molecules, called ribozymes, also exist. Many reactions needed by a biological cell would proceed too slowly under its ordinary conditions without enzymes. See Cooper, The Cell: the central role of enzymes as biological catalysts.
The biological beauty is selective speed. A biological cell contains many possible reactants; its enzymes help determine which transformations happen on a useful timescale. In metabolism, one step's reaction product can become another step's substrate, so selective catalysts support an organized flow of material.
The ribosome is a striking biological example: the machinery that builds proteins has RNA at its catalytic centre. The complete ribosome contains both RNA and proteins, but its bond-forming active site is built from RNA. Structural evidence established this distinction in Nissen et al., The structural basis of ribosome activity in peptide bond synthesis, Science 289, 920–930 (2000).
Selection occurs at an active site, the region where substrate binding and catalytic chemistry take place. Shape and chemical interactions favour some substrates over others; binding can also reshape the enzyme, an induced fit. Recognition alone is insufficient: holding a substrate extremely tightly need not make it react quickly. Catalysis requires a favourable route through the transition state, the high-barrier configuration between reactants and reaction products. Positioning reactive groups and stabilizing this configuration relative to the bound substrate can lower that barrier. Specificity is often strong, but need not mean exactly one acceptable substrate. See Cooper, The Cell: mechanisms of enzymatic catalysis.
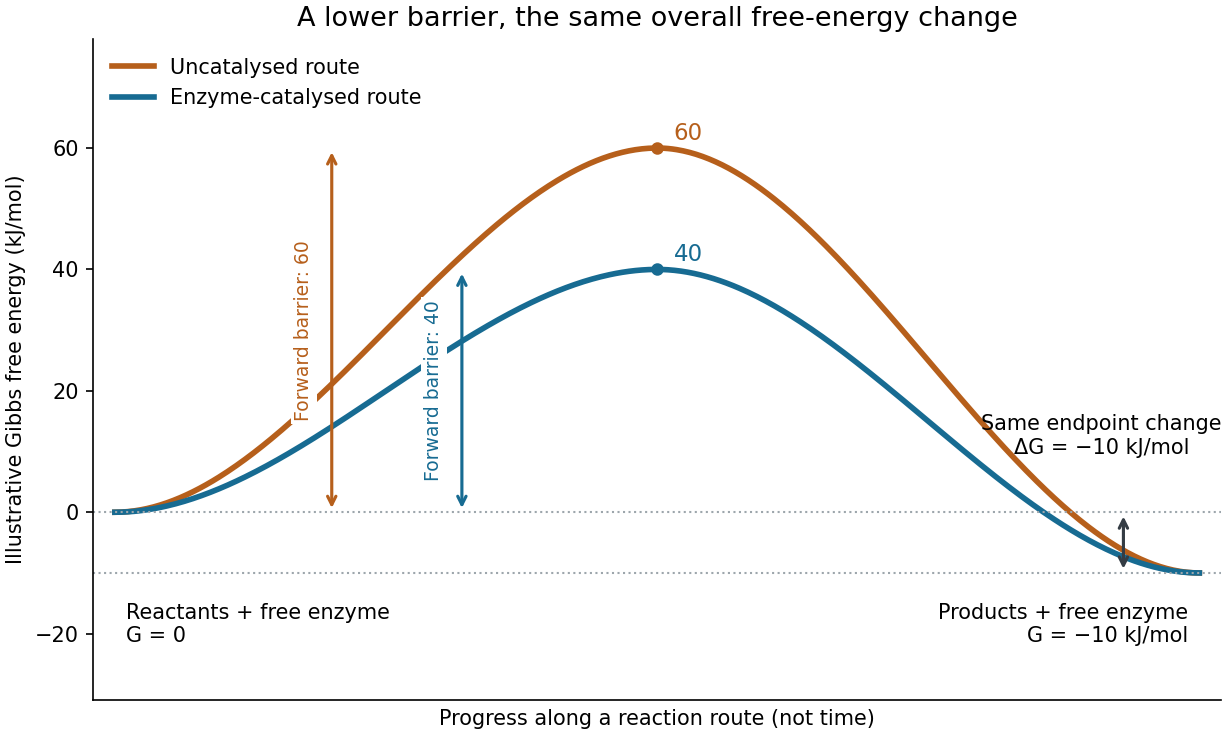
Read the diagram numerically. Its chosen Gibbs free energies are for the reactants and for the reaction products. The uncatalysed peak is , whereas the catalysed peak is . The forward free-energy barriers are therefore and ; viewed from the reaction products, the reverse barriers are and . Both decrease by , while the endpoint difference stays . These are constructed values, not measurements of a particular enzyme.
This separates chemical kinetics from chemical equilibrium. An enzyme changes how rapidly the system approaches chemical equilibrium; it does not change the equilibrium constant for the same overall chemical reaction under the same conditions. Faster catalysis therefore supplies no extra driving energy. In a biological cell, continuing supply of reactants and removal of reaction products can nevertheless sustain a net flow: the surroundings keep that system away from chemical equilibrium. See IUBMB, Symbolism and terminology in enzyme kinetics: basic definitions.
Some enzymes require a cofactor, an additional nonprotein component such as a metal ion or a small organic molecule. Organic helpers are often called coenzymes; they can assist transfers that the protein alone cannot readily perform. The cofactor may remain bound or exchange with the surroundings, depending on the system. A coenzyme can be chemically changed in one reaction and regenerated by another: regeneration of the enzyme does not mean every helper is unchanged in that individual reaction. Some enzymes need no additional cofactor. See Alberts et al., Molecular Biology of the Cell: small molecules associated with proteins and Alberts et al., activated carriers and their regeneration.
How much faster does adding substrate make a reaction? Enzyme kinetics gives a quantitative answer when a model's assumptions fit. For an illustrative reaction obeying Michaelis-Menten kinetics, use the Michaelis-Menten equation with substrate concentration , limiting reaction rate and Michaelis constant . At , substitution gives . At , it gives ; at , it gives . Thus a hundredfold increase in substrate between the first and last examples yields only a tenfold increase in reaction rate.
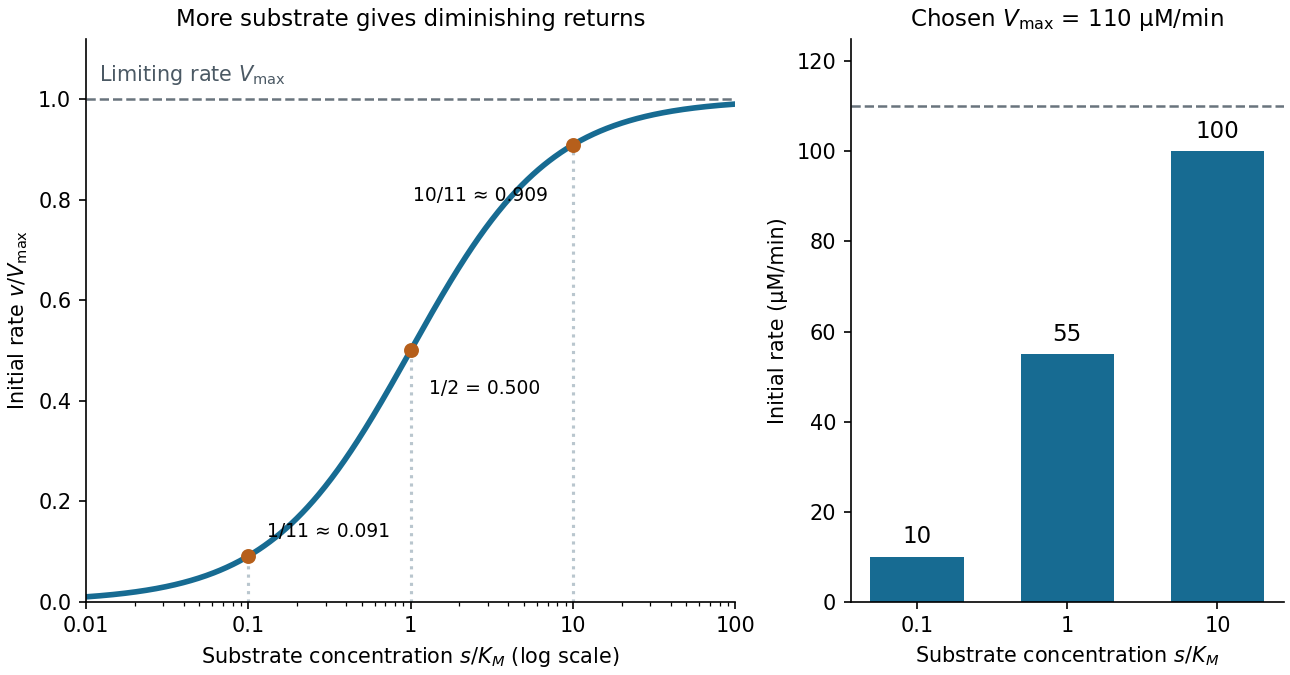
For a chosen , those fractions mean reaction rates of , and . Doubling substrate from to raises the normalized rate only from to , a relative increase of about . In this simplest model, the catalytic machinery is already nearly fully occupied. Doubling the amount of active enzyme instead doubles and the predicted reaction rates at fixed free substrate concentration, provided the other conditions are unchanged. This distinction helps decide whether additional substrate or additional catalytic capacity could improve throughput.
The interpretation has limits. The simplest Michaelis-Menten kinetics assumes a single substrate, fixed active enzyme amount, a suitable quasi-steady-state approximation, and initial measurements with negligible reverse reaction from accumulated reaction product. Substantial depletion, interacting binding sites, or inhibition by excess substrate can invalidate this model. Within it, is the substrate concentration giving half the limiting reaction rate; it is not generally the binding dissociation constant . See IUBMB, enzyme kinetics: steady-state approximation and substrate dissociation and IUBMB, Michaelis-Menten and non-Michaelis-Menten kinetics. Measuring activity also requires specified temperature, pH and necessary cofactor availability: a weak signal can reflect unsuitable conditions rather than little enzyme. See Brooks et al., Assay Guidance Manual: basics of enzymatic assays for HTS.
An enzyme that hydrolyses appropriate glycosidic bonds in starch. Different amylases give different product distributions; pure glucose is not a necessary sole product.
A preparation of milk-clotting enzymes; its action on casein differs from simple acid precipitation.
An enzyme class involved in metabolism of monoamines. Inhibiting it can increase monoamine availability, but that pharmacological fact does not establish a universal deficiency in untreated illness.
An enzyme catalysing ATP hydrolysis. The isolated head of ATP synthase is a rotary ATPase; ATPase activity by itself does not imply proton pumping.
An enzyme transferring electrons from NADPH to oxygen. Plant respiratory-burst oxidase homologues can generate extracellular reactive oxygen species participating in redox signalling.
A peptidoglycan-cleaving enzyme used in controlled cell-wall remodeling and separation; unregulated activity can cause lysis.
A family of bacterial envelope enzymes including peptidoglycan transpeptidases and other activities. The specific catalytic repertoire differs among members.
An enzyme transferring an acyl group to an acceptor; substrate specificity contributes to lipid molecular-species diversity.
The enzyme machinery that assembles fatty-acid chains from activated carbon units through repeated condensation and reduction.
A heme-thiolate enzyme superfamily whose members catalyse diverse oxidative transformations. The name originates from the characteristic approximately 450 nm absorption of the reduced carbon-monoxide-bound form.
A sorghum P450 catalysing conversion of tyrosine toward p-hydroxyphenylacetaldoxime in dhurrin biosynthesis. It has served as a model for photosynthesis-powered heterologous P450 activity.
The highly oxidizing ferryl/porphyrin-radical intermediate used in many P450 oxygenations; its subsequent reactions depend on the substrate and enzyme.
An RNase III enzyme contributing to canonical nuclear microRNA precursor processing. A global processing perturbation affects many RNAs rather than testing one miRNA specifically.
An iron–sulfur enzyme. The cytosolic form associated with IRP1 illustrates switching between enzymatic and RNA-binding functions.
The erythroid aminolevulinate-synthase isoform, with an iron-responsive RNA element coupling its synthesis to iron supply for the haem pathway.
A peptidyl-prolyl isomerase used to study connections between conformational exchange and catalysis.[ref]
Intrinsic dynamics of an enzyme underlies catalysis by Elan Z. Eisenmesser and others. NMR exchange measurements
Adenosine deaminase catalyzes conversion of adenosine and water into inosine and Ammonia. Its zinc-containing active site activates water and stabilizes the reacting configuration; proton transfers assist addition and leaving-group departure. Mutation kinetics constrain catalytic roles but do not uniquely locate every proton.
An enzyme hydrolysing GTP to GDP and phosphate. Many act as nucleotide-controlled switches: hydrolysis changes the state, and nucleotide exchange restores a GTP-bound population for another cycle.
A small GTPase upstream of the budding-yeast mitotic exit network. Its regulation helps couple mitotic exit to spindle position.
An enzyme cleaving a phosphodiester linkage. Cyclic-nucleotide phosphodiesterases reduce cyclic AMP or cyclic GMP signals, so inhibiting them can prolong a messenger response.
An enzyme coupling electrons from reduced ferredoxin to NADP reduction, producing NADPH on the stromal side of the chloroplast electron-transfer system.
An enzyme system hydrolysing glucose-6-phosphate to glucose and phosphate. The hepatic catalytic site faces the endoplasmic-reticulum lumen, so substrate transport is integral to glucose production.
A mitochondrial enzyme converting pyruvate into oxaloacetate using bicarbonate and ATP. It supplies gluconeogenesis and replenishes metabolic intermediates; compartmentation requires suitable subsequent transport routes.
A spatially organized group of three catalytic residues. Serine proteases commonly use serine, histidine and aspartate, but similar geometry in different folds can reflect convergent evolution.
An enzyme hydrolysing beta-galactosides. In the lac operon system, its processing of lactose includes production of allolactose, which regulates LacI.
In mechanism-based inhibition, catalytic processing of an inhibitor generates the species responsible for sustained inhibition. The covalent bond can be between inhibitor and cofactor rather than to an amino acid side chain; persistence must be described on the relevant timescale.
Steroid 5α-reductase reduces the conjugated double bond of particular steroids using NADPH. Human type 2 can convert testosterone to dihydrotestosterone and generate an inhibitory cofactor adduct from finasteride.
An enzyme catalysing decomposition of hydrogen peroxide into water and oxygen. Compartment-specific activity can be used to test contributions of peroxide buffering.
A family of proteolytic enzymes involved in processes including apoptosis. A caspase reporter is evidence about pathway activity and should be interpreted together with cellular morphology and subsequent fate.
AceK is a bifunctional isocitrate-dehydrogenase kinase/phosphatase. Metabolic effectors regulate its competing activities; in Pseudomonas aeruginosa it controls ICD but not the separate IDH enzyme.
malate synthase combines glyoxylate with an acetyl unit from acetyl-CoA to form malate. Together with isocitrate lyase it provides the characteristic carbon-conserving route of the glyoxylate shunt.
isocitrate lyase cleaves isocitrate into succinate and glyoxylate. It competes with isocitrate dehydrogenases for the branchpoint substrate and initiates the glyoxylate shunt.
In the Pseudomonas aeruginosa branchpoint discussed here, IDH denotes the isocitrate dehydrogenase activity insensitive to AceK phosphorylation and regulated allosterically by metabolic effectors. The abbreviation is context-dependent across organisms.
A cytosolic isocitrate-dehydrogenase isoform. Certain cancer-associated variants acquire an altered catalytic activity producing 2-hydroxyglutarate.[ref]
Cancer-associated IDH1 mutations produce 2-hydroxyglutarate by Lenny Dang and others. Biochemical characterization of mutant enzyme
In the Pseudomonas aeruginosa branchpoint discussed here, ICD is the isocitrate dehydrogenase susceptible to AceK-dependent phosphorylation. It must be distinguished from the separate IDH activity, which is not regulated by that phosphorylation route.
A cellulase contributes to cellulose hydrolysis. Activity on a soluble cellulose derivative need not predict activity on an ordered microfibril or a lignified wall.
A lytic polysaccharide monooxygenase is a copper-dependent enzyme that cleaves polysaccharides oxidatively. Its performance depends on substrate access, reduction and oxidant supply; uncontrolled peroxide can damage the catalyst.
A beta-glucosidase hydrolyzes beta-glucosidic substrates. Conversion of cellobiose to glucose can reduce product accumulation during cellulose saccharification.
A cellobiohydrolase releases cellobiose units from glucan chains, often through processive cleavage at an accessible chain end. Chain binding and extraction from an ordered substrate can limit activity.
An endoglucanase cleaves internal bonds in an accessible glucan chain. On cellulose it can generate new chain ends for complementary processive enzymes.
A fatty acid elongase participates in acyl-chain extension. An enzyme acting on an acyl-CoA substrate may not directly access a chain held in a phospholipid.
A fatty acid desaturase introduces a double bond into an acyl substrate. Chain length, double-bond position, carrier preference and electron-supply compatibility matter when transferring the activity between organisms.
cellulose synthase polymerizes glucose into cellulose using an activated donor. Membrane-associated systems can couple chain elongation to extrusion through a membrane channel.
PglB is a bacterial oligosaccharyltransferase whose structural characterization supplies a model for protein-acceptor recognition. Its donor and complete pathway should not be assumed identical to those of eukaryotic ER glycosylation.
oligosaccharyltransferase transfers an assembled oligosaccharide from a lipid-linked donor to a protein acceptor. Recognition of an accessible acceptor sequence is distinct from synthesis of the donor.
A glycosyltransferase transfers a glycosyl group from a donor to an acceptor. Donor recognition, acceptor position and stereochemical outcome distinguish different enzymes.
lysyl oxidase contributes to covalent cross-link formation in extracellular matrix proteins. Perturbing its activity can change matrix mechanics and chemistry together, which matters when interpreting mechanotransduction experiments.
PI3K-C2α is a class II phosphoinositide 3-kinase implicated in local phosphoinositide production during endocytosis. Catalytic rescue and lipid measurements help distinguish its enzymatic role from other consequences of its depletion.
synaptojanin is a phosphoinositide phosphatase involved in endocytic lipid turnover. Loss of its activity can affect coat removal and synaptic vesicle recycling; genetic defects need to be separated from secondary developmental changes.
A luminescent reporter enzyme whose output can report promoter activity under controlled expression and assay conditions.
A luminescent reporter enzyme often used as a separately measured reference in a dual-luciferase assay. Valid normalization requires a reference insensitive to the experimental manipulation.
An enzyme removing a phosphate group from an appropriate substrate. In a phosphorylation assay, inhibition preserves the starting state while controlled phosphatase treatment can test phospho-dependent detection.
An enzyme that hydrolyzes the beta-lactam ring of susceptible antibiotics. Substrate specificity and catalytic mechanisms vary among classes and variants.
A TEM-family beta-lactamase variant whose structure and substrate specificity differ from TEM-1. Controlled comparisons distinguish access remodeling, catalytic behavior and altered expression.[ref]
A reference TEM-family class A beta-lactamase against which altered-spectrum variants can be compared.
An enzyme catalyzing reactions involving molecular hydrogen, including disposal of reducing equivalents in certain anaerobic metabolic pathways.
An enzyme interconverting ATP, ADP and AMP through . Near equilibrium, AMP:ATP is proportional to the square of ADP:ATP under fixed conditions.
A covalent enzyme intermediate bearing an acyl group during catalysis. Its formation and breakdown can occur at different rates, producing burst kinetics if the first released reaction product reports a fast step.
An enzyme that adds adenosine residues to an RNA end using ATP. Supplying a correctly pre-cleaved substrate separates the extension reaction from the preceding cleavage step.
STN7 kinase is required for plant LHCII phosphorylation associated with state transitions. Mutant and complementation experiments can separate its regulatory role from general loss of photosynthetic competence.
Violaxanthin de-epoxidase catalyses steps of conversion from violaxanthin towards zeaxanthin under suitable thylakoid-lumen conditions. Its activity links lumen acidification to a change in antenna pigment composition.
Creatine kinase catalyses reversible transfer between phosphocreatine and ADP to yield creatine and ATP. Inferring free ADP from this reaction requires an appropriate equilibrium model and information about pH, magnesium and the creatine pool.
Glycogen synthase extends glycogen using activated glucose donors. Its activity is regulated by phosphorylation and metabolites, and its activation can be coordinated with inactivation of glycogen phosphorylase.
Protein phosphatase 1 catalyses protein dephosphorylation in regulated complexes. Association with targeting partners matters: effects on one substrate do not imply indiscriminate dephosphorylation of all cellular proteins.
A Kemp eliminase is a catalyst for Kemp elimination. Computationally designed protein examples allow tests of active-site construction and subsequent optimization by directed evolution.
Subtilisin E is a protease used as a model in protein-engineering experiments. Sequential mutagenesis and screening for activity in organic solvent illustrate optimization without requiring a complete predictive model of every substitution.
A histone acetyltransferase catalyses acetyl-group transfer to histone residues. The resulting changes in charge and reader-protein recruitment can influence chromatin accessibility; the effect depends on the modified site and regulatory context.
An enzyme that separates DNA strands using chemical energy, enabling replication and other DNA transactions.
A reverse-transcriptase-containing enzyme that extends telomeric DNA using an associated RNA template, contributing to chromosome-end maintenance in many eukaryotes.
Flap endonuclease 1, involved in processing displaced DNA structures during eukaryotic lagging-strand maturation and other DNA transactions.
A topoisomerase that transiently breaks both strands of one DNA duplex to permit another segment to pass, contributing to resolution of replicated chromosome interlinks.
An enzyme that changes DNA topology through controlled strand breakage and rejoining, helping resolve supercoiling and molecular interlinking.
An enzyme attaching an appropriate amino acid to a cognate transfer RNA. Reduced charging can slow decoding even if ribosome initiation remains active.
Correction of inappropriate amino-acid activation or tRNA charging. Correct codon recognition cannot repair a tRNA charged with the wrong amino acid.
An enzyme that removes acetyl groups from histones. Its local recruitment can oppose histone acetylation and favor repressive chromatin.
An enzyme that adds methyl groups to specified histone residues. Its presence alone does not specify repression, because distinct modified sites can have different regulatory roles.
A histone methyltransferase contributing to repressive H3K9 methylation, including at targets associated with KAP1.
The fission-yeast histone methyltransferase responsible for H3K9 methylation. Its chromodomain couples recognition of an existing mark to further writing.
An enzyme that changes nucleic-acid topology by transient strand cleavage and resealing. Activity on DNA can change DNA supercoiling.
A nuclease that cuts internal bonds of a nucleic-acid chain. For an RNA substrate, internal cleavage can create new ends for further degradation.
A recombinase is an enzyme that catalyzes a genetic recombination reaction. The RecA family performs homology-dependent DNA strand exchange; the term also includes enzymes with other recombination mechanisms.
dihydrofolate reductase catalyzes reduction of dihydrofolate to tetrahydrofolate using NADPH. Its catalytic cycle is a model for connecting protein dynamics, binding and release.[ref]
The dynamic energy landscape of dihydrofolate reductase catalysis by David D. Boehr and others. pp. 1638–1642, dynamic states across the catalytic cycle
A nuclease is an enzyme that cleaves nucleic acids. ribonucleases act on RNA; other activities act on DNA. Nuclease sensitivity can reveal a nucleic-acid-dependent assembly, but loss of association can also reflect secondary structural disruption.
A nuclease participating in primary piRNA processing in Drosophila, distinct from the Dicer cleavage pathway.
An enzyme that processes appropriate double-stranded RNA substrates into small guide-RNA precursors in several RNA interference pathways.
Articles were limited to the first 100 out of 192 total.
Articles by others on the same topic



For an initial concrete example, consider e. Coli K-12 MG1655 gene thrA.
How Enzymes Work by RCSBProteinDataBank (2017)
Source. Shows in detail how aconitase catalyses the citrate to isocitrate reaction in the citric acid cycle.