A catalyst makes a chemical reaction proceed faster by participating in an alternative reaction mechanism and being regenerated. This makes repeated conversion possible: the same catalyst can help many successive reactants become reaction products. An enzyme is a biological catalyst. The overall standard change in Gibbs free energy is unchanged; see the IUPAC Gold Book: catalyst.
Regeneration does not mean that the catalyst sits unchanged throughout every step. For example, a schematic reaction mechanism can temporarily bind A to a catalyst C in a reaction intermediate AC:Adding the steps cancels both the reaction intermediate AC and the catalyst C from the net chemical equation. AC is produced and later consumed; C is used and later recovered. These letters describe possible roles, not a measured reaction mechanism. The alternative route is useful only if its chemical kinetics permits faster conversion. See Flowers et al., Chemistry 2e, §12.7: catalysts and alternative mechanisms.
For a worked barrier example, consider an idealized reversible A-to-B conversion at fixed temperature . Stipulate standard molar Gibbs free energy levels and , with effective transition state levels without a catalyst and with a catalyst. Use the same standard states and the same zero for both routes; the catalytic route includes the regenerated C at both endpoints. Unlike the separate potential-energy profile example, this illustration uses Gibbs free energy at one specified temperature.
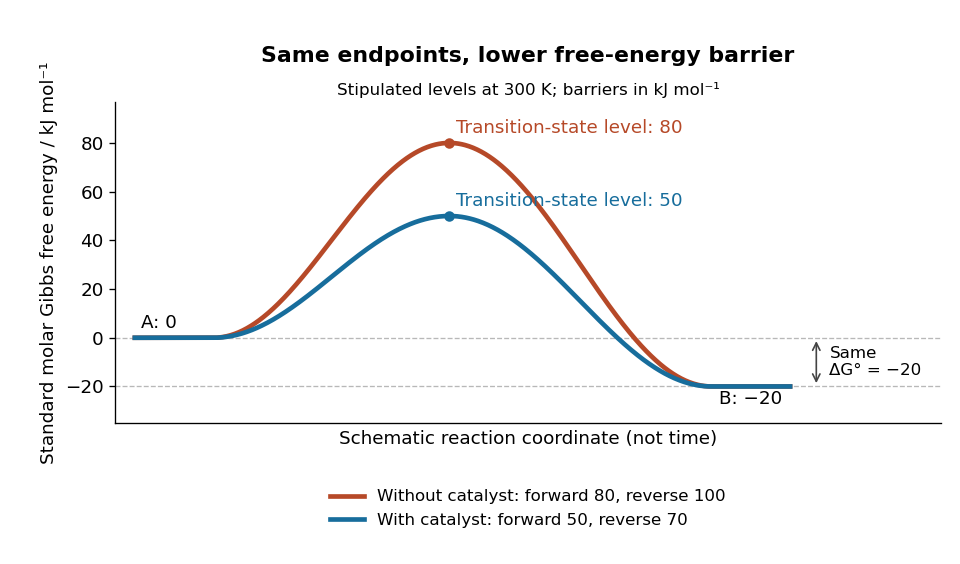
A catalyst lowers the stipulated Gibbs energy of activation in both directions while retaining the same endpoints
. Original smooth schematic Gibbs free energy profiles at , with levels in . The horizontal reaction coordinate describes progress along each schematic route, not time. These are chosen effective barriers, not measured potential energy surfaces, actual molecular paths, or the two-step reaction mechanism above. A real catalytic route can have several transition states and reaction intermediates. Details.
The Gibbs energy of activation is measured from the starting endpoint in the chosen direction. Thus the four barriers areBoth directional barriers fall by . The endpoint difference stays .
To turn those barriers into rate constants, use the Eyring form of transition state theory for this effective first-order reaction pair, with a common prefactor and unit probability of successful passage:Here is the Boltzmann constant, is the Planck constant, and is the molar gas constant. The common prefactor cancels from the comparisons. These Gibbs energies of activation are not the empirical activation energy of the Arrhenius equation. With the stipulated value , the speedup isThe exponent is a dimensionless quantity: convert to before dividing by . The exponential function explains why a modest barrier reduction can have a dramatic kinetic effect. This number follows from the chosen free-energy barriers and common prefactor; it is not a universal catalytic multiplier or a prediction across temperatures.
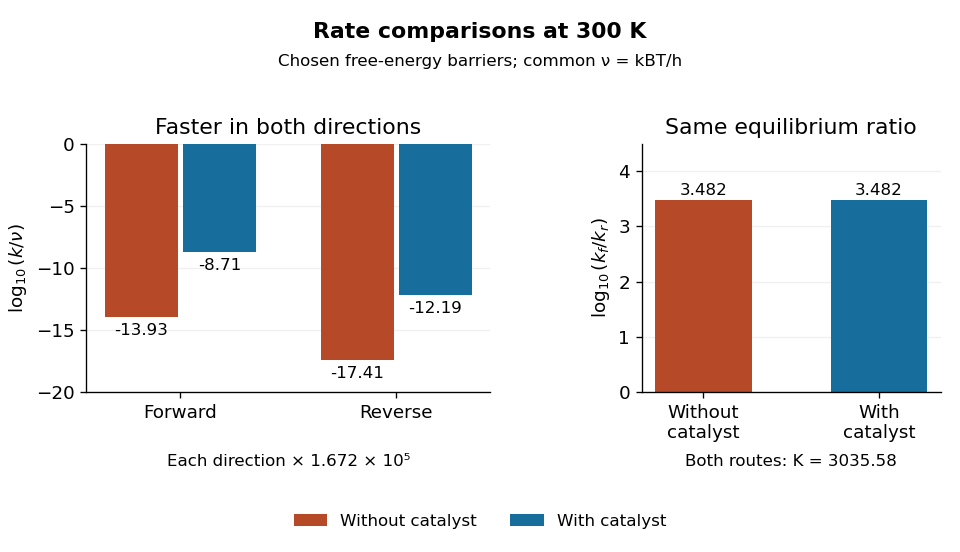
Both directional rate constants increase by about , while their forward-to-reverse ratio stays about
. Original bar plots at . Left: base-ten logarithms of the dimensionless normalized rate constants for the four stipulated barriers. A less negative value means faster conversion. Right: the logarithm of is identical for the two routes. Absolute units factor out through the common ; no temperature dependence outside this one model point is asserted. Details.
Assume ideal thermodynamic activities with a common concentration standard state, and directional rate laws and . At chemical equilibrium, equal reaction rates give for this model. The barriers giveThe catalyst speeds up both directions, so it changes how quickly chemical equilibrium is approached without changing this equilibrium constant. For a general chemical reaction, the thermodynamic relation is with consistent standard states; an arbitrary multistep reaction mechanism need not permit for two measured rate constants. At the same temperature, pressure, and thermodynamic activities, a catalyst also leaves the actual overall reaction change in Gibbs free energy unchanged. See 13.3 Shifting Equilibria: Le Châtelier’s Principle (§13.3: a catalyst does not shift equilibrium). To drive an unfavourable overall conversion, a process such as energy coupling must change the combined transformation.
A supplied precursor that must be chemically converted into the catalytically active form. Activation stoichiometry need not equal the reagent consumption of each subsequent catalytic turnover.
Transition state theory relates an elementary reaction's rate constant to a Gibbs energy of activation. The Eyring expression used in the catalyst example assumes every passage counted at the transition state succeeds. More generally, a factor accounting for unsuccessful passage multiplies it. Comparing barriers alone gives the quoted speedup only when the remaining prefactors agree. Free-energy profiles can depend on temperature, so the example's values at do not determine a temperature curve. See the IUPAC Gold Book: transition state theory.
A constant tensile force changes a one-coordinate free-energy landscape from to . To first order, an escape barrier decreases by , where is the saddle-to-initial-minimum displacement at zero force. An approximately fixed attempt prefactor gives the displayed rate law. Its characteristic force is . Large changes in saddle location or competing pathways invalidate a constant activation distance. The barrier is measured relative to the initial state; an absolute saddle energy is not enough to determine an escape rate.
The pressure derivative of the standard Gibbs energy of activation: . In the corresponding rate convention, . A negative value makes increasing pressure accelerate the reaction. Solvent reorganization can contribute as well as the intrinsic reacting geometry.
The standard entropy difference between the activated-complex reference state and the reactants. Its numerical value depends on standard-state conventions; association, molecular reorientation, internal motions and solvent reorganization can all contribute.
The standard enthalpy difference between the activated-complex reference state and the reactants at the same temperature and pressure. Under the usual Eyring assumptions it controls the slope of against .
A transition state theory expression relating a rate constant to the standard free energy of activation. For a bimolecular reaction expressed in concentration units, in the unit-transmission approximation.
The Gibbs energy of activation is the standard molar Gibbs free energy change associated with going from the starting reactants to the transition state in transition state theory. It specifies the directional barrier, rather than the overall endpoint change in Gibbs free energy. Use consistent standard states when comparing Gibbs energies of activation and rate constants. See the IUPAC Gold Book: Gibbs energy of activation.
The standard molar enthalpy difference between the transition state and the starting reactants, at the same temperature and pressure, in transition state theory.[ref]
The standard molar entropy difference between the transition state and the starting reactants, at the same temperature and pressure, with specified standard states, in transition state theory.[ref]
Articles by others on the same topic
There are currently no matching articles.