The nine normal coordinates transform as . Place the molecule in the plane, with its four outer atoms on the and axes, as specified for the vertical reflection planes. In the class orderthe fifteen Cartesian displacements have characters of a representationFor each operation this character is the number of fixed atoms times the trace of its polar-vector matrix. Subtract translations and rotations , givingMolecular character reduction in the D4h molecular point group yields the stated decomposition. Its dimensions sum to , providing an independent degree-of-freedom check.
An infrared fundamental must transform like a component of the electric dipole moment; a Raman spectroscopy fundamental must transform like a component of the symmetric polarizability tensor. For this centrosymmetric D4h molecular point group, the resulting classification is:
Here parallel and perpendicular refer to the principal fourfold axis. The table obeys infrared and Raman mutual exclusion.
Three separated infrared fundamentals: one parallel band and two perpendicular bands. Each frequency belongs to a doubly degenerate mode, but degeneracy does not create two distinct band origins.
The square-planar rotor is an oblate symmetric top. A parallel band has and , producing P and R progressions with approximate spacing and a central Q contribution from ; the parallel Q branch is absent. At modest resolution many nearly coincident components merge.
A perpendicular band has and . Its Q subbranches produce a characteristic series of sharper features on the broader unresolved P/R envelope. Neglecting changes of rotational constants on vibration, successive perpendicular Q features are separated by approximately . These are rotational contours rather than extra normal-mode fundamentals.
All the listed modes are Raman active; only and are infrared active. For the C4v molecular point group, with vertical planes through the outer atoms:
| Symmetry | Number of modes of this symmetry | Infrared | Raman | 2 | Parallel, | Yes: | 2 | No | Yes: | 1 | No | Yes: | 2, each doubly degenerate | Perpendicular, | Yes: |
|---|
There is no inversion centre, so infrared and Raman mutual exclusion does not apply.
In the stipulated ideal, fully resolved limit, yes: square-planar geometry gives three infrared fundamentals, whereas square-pyramidal geometry gives four: two parallel bands and two perpendicular bands. The extra parallel fundamental is the distinguishing feature. In a real incomplete spectrum, a weak band, overlap or accidental degeneracy could hide that difference, so absence of a fourth observed band would not alone prove planarity.
. The parallel-band progression gives , so . The perpendicular Q-feature spacing givesFor an oblate symmetric top, , hence and . The rotational constant relation then givesThe estimated constants neglect centrifugal distortion and the change of rotational constants in the excited vibration.
; the central atom makes no contribution. Let be the radius of the square of outer atoms and the central atom's height above its plane. Then . Adjacent bond vectors have perpendicular in-plane components, soThe centre of mass lies on the fourfold axis, even when . Thus the central atom and the shift of origin along the axis contribute no perpendicular lever arm. The four outer atoms each contribute , givingThe result concerns the smallest adjacent bond angle; the angle between opposite outer atoms is a different angle in a pyramid.
With integer fluorine mass and ,This angle is below the planar limit , consistent with the proposed pyramid. The sulfur mass does not enter this axial moment of inertia.
The inferred dipole moment is approximately . With the static field parallel to the radiation electric field, the microwave selection rule is . A fully resolved three-component first-order rotational Stark effect pattern naturally identifies the , transition: the three initial values give the three components. would have no first-order shift.
Subtract the lower-state field shift from the upper-state field shift, rather than equating a transition shift to one level's shift:For and , neighbouring components are separated by . HenceThis assignment assumes the stated splitting resolves all allowed first-order components, rather than three accidentally unresolved groups of a higher- transition.
For the absorption spectrum, pass a scanned monochromatic beam through the solution and a matched solvent reference. Record the incident and transmitted intensities and calculate the absorbance , using an appropriate blank and concentration to avoid saturation.
For the fluorescence emission spectrum, excite at a fixed wavelength within an absorption band, collect emission away from the incident beam, commonly at , and scan an emission monochromator. Correct the detected signal for wavelength-dependent detector sensitivity, subtract solvent/background light and exclude scattered excitation light. The two plotted vertical scales have been independently normalized; their equal peak heights do not imply equal absorption and emission rates or unit fluorescence quantum yield.
Vibrational and solvent relaxation remove energy before emission, producing a Stokes shift. Absorption initially excites a vertical transition from the populated ground-state vibrational levels to excited-state vibrational levels. Vibrational relaxation and solvent rearrangement then bring the molecule near the relaxed lowest excited singlet level before fluorescence. Emission ends in vibrationally excited levels of the electronic ground state, so the emitted photon generally has less energy and a longer wavelength. This explanation does not require a change of electronic multiplicity.
The approximate mirror relationship follows from the Franck-Condon principle when the ground and excited electronic states have similar vibrational frequencies and comparable potential shapes displaced along the active coordinates. Absorption samples overlaps, while relaxed emission samples overlaps. For displaced harmonic potentials, the corresponding overlap intensities form similar progressions on opposite sides of the transition between the lowest vibrational levels of the two electronic states.
The symmetry is approximate and is most naturally compared on an energy or wavenumber axis. A wavelength axis is nonlinear in photon energy, and solvent response, frequency changes, differing transition strengths and instrumental corrections can spoil an exact mirror image.
For a fluorescence excitation spectrum, hold the emission detector at a fixed wavelength within the fluorescence band and scan the excitation wavelength. Correct for the incident photon flux and instrumental response at each excitation wavelength.
For one emitting molecular species with an approximately excitation-independent fluorescence yield, it should follow the absorption spectrum. In a dilute sample the detected emission is proportional to the photons absorbed, . Without the dilute approximation it follows absorptance rather than absorbance exactly. Impurities, photochemistry or multiple emitting states could invalidate that correspondence.
The phosphorescence quantum yield is the number of phosphorescence photons emitted per photon absorbed. In rate form,It is dimensionless. The denominator counts absorbed photons, not incident photons or the instantaneous triplet population.
Almost every absorbed photon generates a triplet, within the stated kinetic scheme. The branching fraction from the excited singlet isso under steady illumination. This statement concerns the triplet formation yield; it does not mean that the triplet population or the phosphorescence yield is also one. The triplet can subsequently deactivate or be quenched.
The steady-state approximation for the excited singlet givesand that for the triplet givesCombining these with the quantum yield definition gives the more general expressionUnder the supplied limit , this becomesThe approximation is the near-unit triplet formation yield, not an assumption that phosphorescence is the only triplet loss process.
Without quencher, . Taking the ratio gives the Stern-Volmer equationThus the plot has unit intercept and slope . A triplet formation yield less than one would cancel from this ratio if it remained independent of quencher concentration. The result assumes dynamic triplet quenching and no additional quencher-induced absorption or singlet loss.[ref]
Stern-Volmer kinetic relationships in the IUPAC Gold Book. Definition and dynamic-quenching relation between quantum-yield ratios, quencher concentration and unquenched lifetime
. The unquenched triplet lifetime isSince the slope is , the bimolecular rate constant isFor comparison, the unquenched phosphorescence yield in the unit-triplet-yield approximation is .
The measured decay is approximately consistent with the predicted loss rate. Following rapid singlet-to-triplet population transfer, the unquenched phosphorescence intensity obeysA straight-line fit of to the four times, expressed in seconds, givesThis differs from by about . For example, using the first point and the predicted rate gives subsequent intensities about , close to the supplied . The data support the expected millisecond exponential decay to this precision. With no measurement uncertainties supplied, a formal statistical acceptance or rejection is not justified.
Let denote a small longitudinal displacement from equilibrium, and let every atomic mass be . The constant equilibrium lengths disappear from the second derivatives. The force constant matrix isThe mass-weighted dynamical matrix is , with equations . If the phrase dynamical matrix denotes the unweighted Hessian, that object is instead; the normal-mode equation is in either notation. Every row sums to zero, as required for a freely translating chain.
At , each independent diatomic spring has one translation and one stretch. Consequently the angular frequencies areThe two zero modes are independent centre-of-mass translations of the two molecules. There are no transverse coordinates in this model; adding transverse degrees of freedom would be a different problem.
For the collinear centrosymmetric arrangement, the longitudinal coordinates transform in the D infinity h molecular point group. Inversion sends , because displacement is a polar vector. ThereforeAn orthonormal set of symmetry coordinates is:
| Coordinate | Displacement vector | Symmetry | at |
|---|
Here translates the complete system, separates the molecular centres, stretches both molecules together, and lengthens one while shortening the other. Reversing every arrow changes the coordinate sign, not its physical mode.
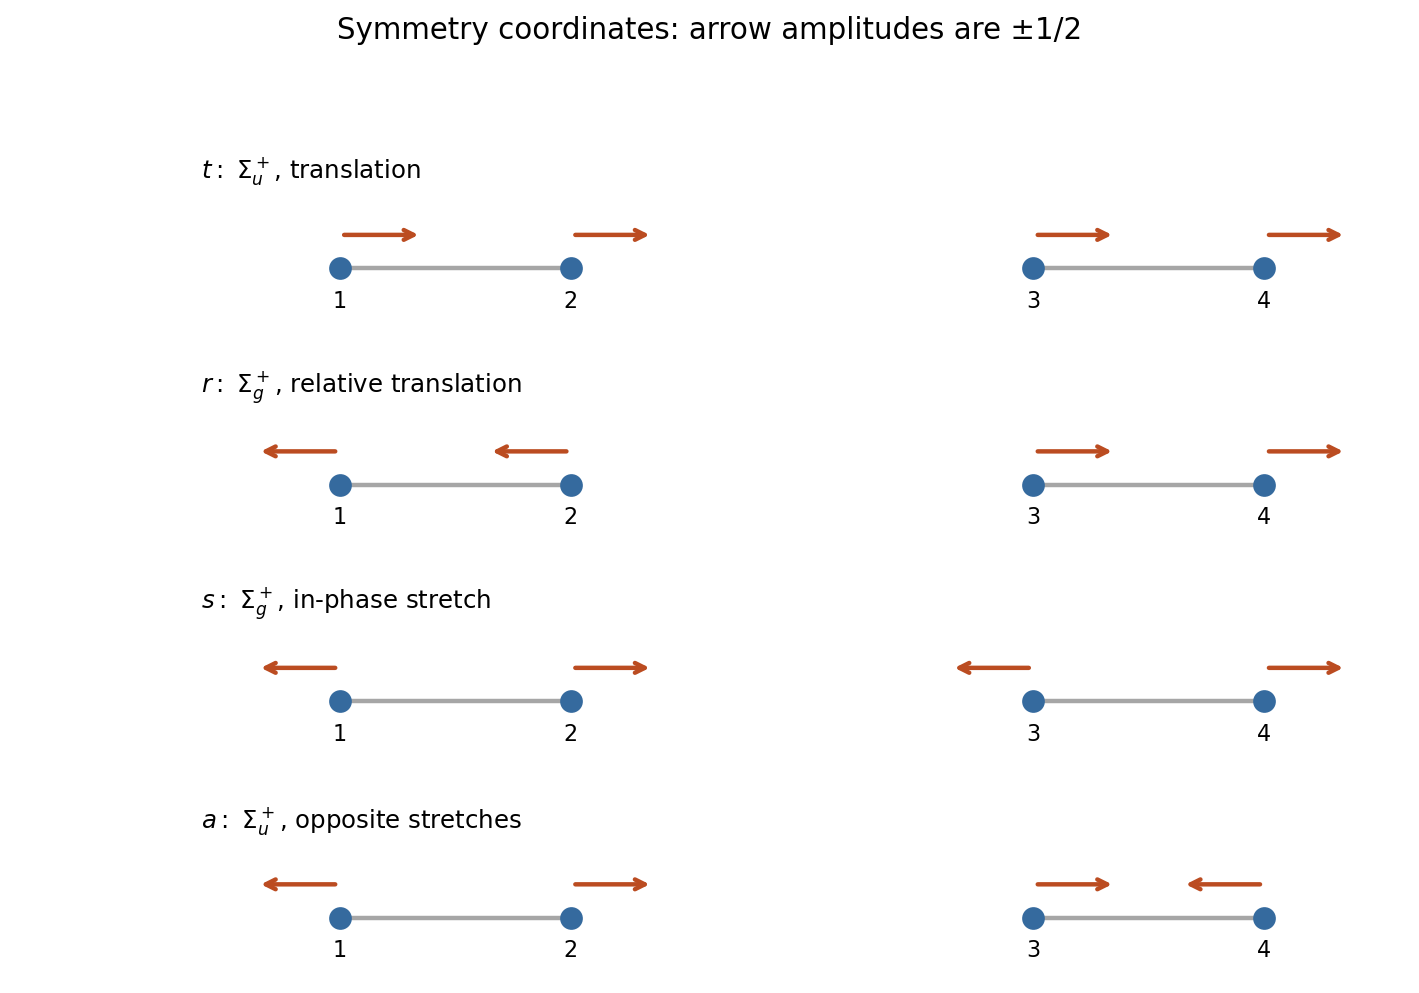
The nontotally symmetric longitudinal modes are the two coordinates and . Both have , so the middle spring is not extended. Hence its spring constant cannot alter their energies:These results are exact for every positive , not just perturbative. The zero mode expresses overall translation invariance.
In the totally symmetric basis, the force constant matrix reduces toThe unperturbed eigenvalues are distinct within this symmetry block, so nondegenerate perturbation theory applies even though the full unperturbed matrix has degeneracies in different symmetry sectors. The first-order shifts are both . At second order the off-diagonal matrix element gives for the lower mode and for the upper one. ThereforeIn frequency form,The corresponding eigenvectors are and . As a check, the exact squared-frequency eigenvalues arewhose expansions reproduce the perturbation theory result. The newly lifted zero frequency scales as , so expanding as an ordinary Taylor series in would be inappropriate.
and . With and , the dilute-gas stretch givesUse in here. The shifted line is the higher totally symmetric mode; the other molecular stretch remains at . To first order,The ratio validates the weak-coupling expansion. Inverting the exact eigenvalue instead, with , givesa negligible change at the precision of the model.
For clamped nuclei and SI units, the many-electron electronic Hamiltonian isThe terms are electron kinetic energy, electron–nucleus attraction, and electron–electron repulsion. The nuclear kinetic operator is omitted in the Born-Oppenheimer approximation. The nuclear repulsionis a constant at fixed separation and is added to electronic energies to form the total potential curve. Some conventions include it in as a constant; the electronic-energy convention used below excludes it so that the two one-electron energies can be compared without double-counting nuclear repulsion.
An effective one-electron Hückel method operator may be writtenThe first term is one-electron kinetic energy. The effective site potentials represent attraction to the nuclei and any averaged screening implicit in the chosen parametrization. They replace the explicit many-electron interaction; they do not contain a coordinate-dependent two-electron operator .
For normalized hydrogen-centred orbitals, symmetry givesThe Hückel Coulomb parameter is an on-site one-electron energy, not the electron–electron Coulomb repulsion integral. The Hückel resonance integral couples the sites; with the usual same-phase atomic-orbital convention it is negative and stabilizes the bonding combination. For zero overlap, the two energies are .
A doubly occupied spatial orbital gives a spin-singlet Slater determinant:Equivalently, . The spatial factor is symmetric, and the normalized antisymmetric spin factor enforces the Pauli exclusion principle. Here includes position and spin.
Let be real, with each atomic orbital normalized. ThenTogether with orthonormal spin functions, this also normalizes the Slater determinant: its norm squared is . Under the subsequent zero-overlap assumption,using the specified nonnegative branch for .
The physical curve has its bonding minimum near , with ionic endpoints at . Use electronic energies excluding the fixed . Define , , and . The one-electron contribution isThe Coulomb energy of the doubly occupied orbital isso . For localized orthogonal orbitals, a transparent illustrative approximation neglects integrals involving the product and retains the on-site repulsion and intersite direct repulsion :Since normally , spreading the occupied orbital over both atoms reduces the repulsion. At equilibrium the bonding one-electron term favours , so the minimum represents the Restricted Hartree-Fock method approximation to . At or , both electrons occupy one atom, giving the same ionic energy . At the orbital is antibonding and the one-electron contribution is raised.
The diagram below uses an illustrative set of integrals to show these landmarks; it is not a numerical calculation for the molecule. Zero orbital overlap alone does not force every exchange or three-index Coulomb integral to vanish, so the simplified expression is an explicit sketching approximation, not a consequence of orthogonality.
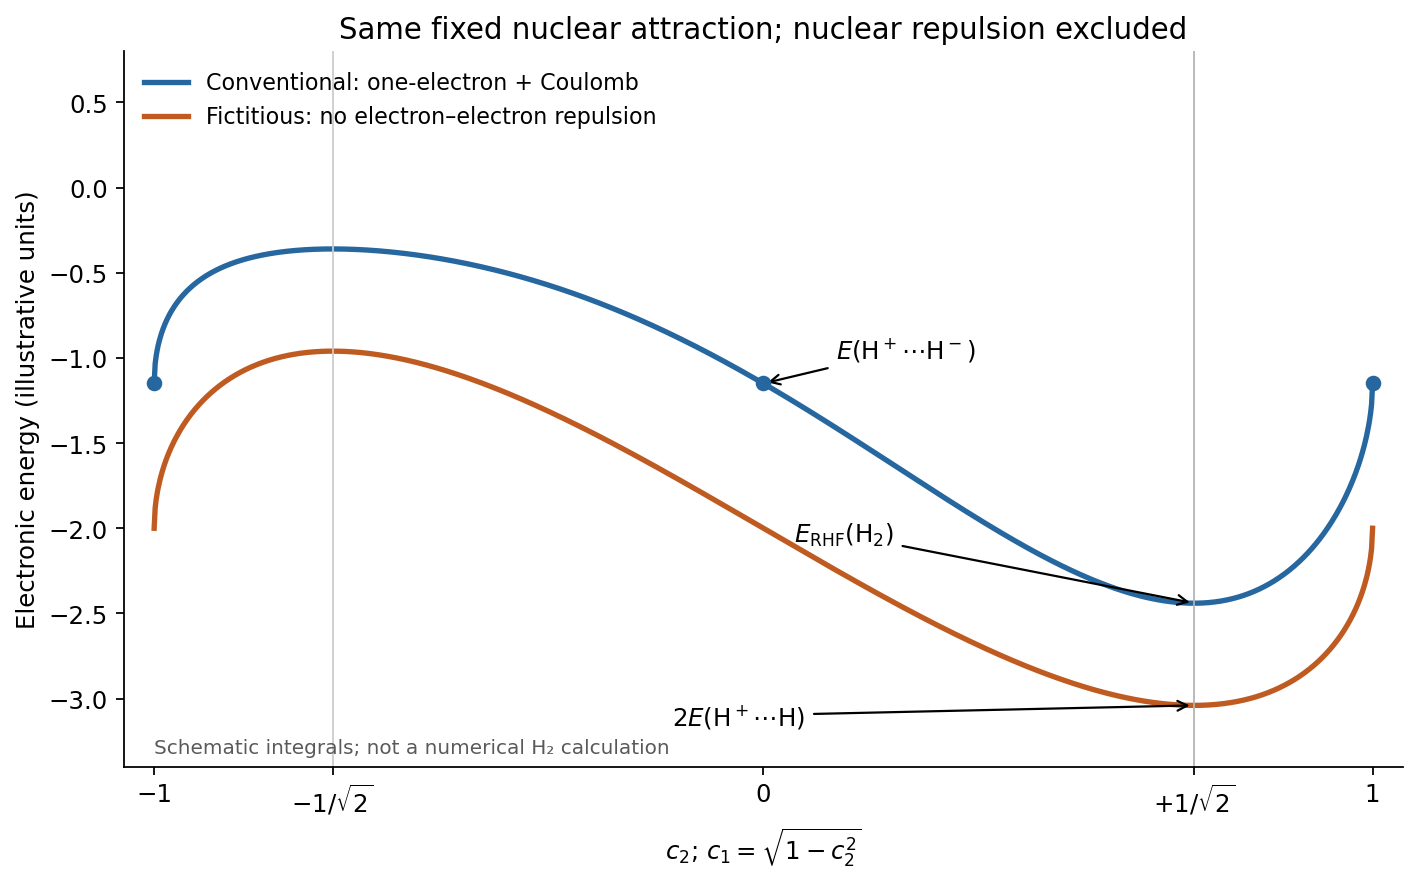
Removing electron–electron repulsion lowers the entire curve and exaggerates the bonding stabilization. In the same fixed-orbital notation and electronic-energy convention,Its minima and maxima occur at and , respectively; all three localized points have energy . The physical curve lies above it by the positive Coulomb expectation .
For the one-electron problem, the bonding energy is . Two noninteracting opposite-spin electrons may both occupy that orbital, so the lower curve's bonding minimum isThis is the third requested energy label in the common graph. If total molecular energies include nuclear repulsion, the corresponding identity is , because the two-electron molecule contains only one nuclear pair. An effective one-electron potential re-fitted after removing repulsion would be a different comparison; here the same kinetic and nuclear-attraction operators are retained.
Restricted Hartree–Fock incorrectly retains ionic configurations when a homonuclear bond dissociates. As overlap vanishes, its symmetry-preserving occupied orbital approaches . The doubly occupied spatial product containsThe first and last terms place both electrons on one separated atom. Their combined probability remains one half, while the correct neutral singlet has one electron on each atom. The resulting spurious Coulomb repulsion raises the restricted dissociation limit. This is missing static electron correlation, not a failure of the Born-Oppenheimer approximation.
A two-configuration expansion using both and can cancel the ionic terms and recover the Heitler–London separated-atom singlet. Configuration interaction in the appropriate active space or a multiconfigurational self-consistent treatment supplies this remedy. The Unrestricted Hartree-Fock method can instead localize opposite spins on different atoms and give the correct separated-atom energy, but its single determinant breaks spin symmetry and is not the pure singlet at intermediate separation. The distinction between restricted and symmetry-broken descriptions of stretched hydrogen is treated in the original analysis of Coulson and Fischer.[ref]
XXXIV. Notes on the molecular orbital treatment of the hydrogen molecule by Charles Alfred Coulson and Inga Fischer-Hjalmars. Philosophical Magazine 40 (1949), 386–393, molecular-orbital treatment of hydrogen and its dissociation
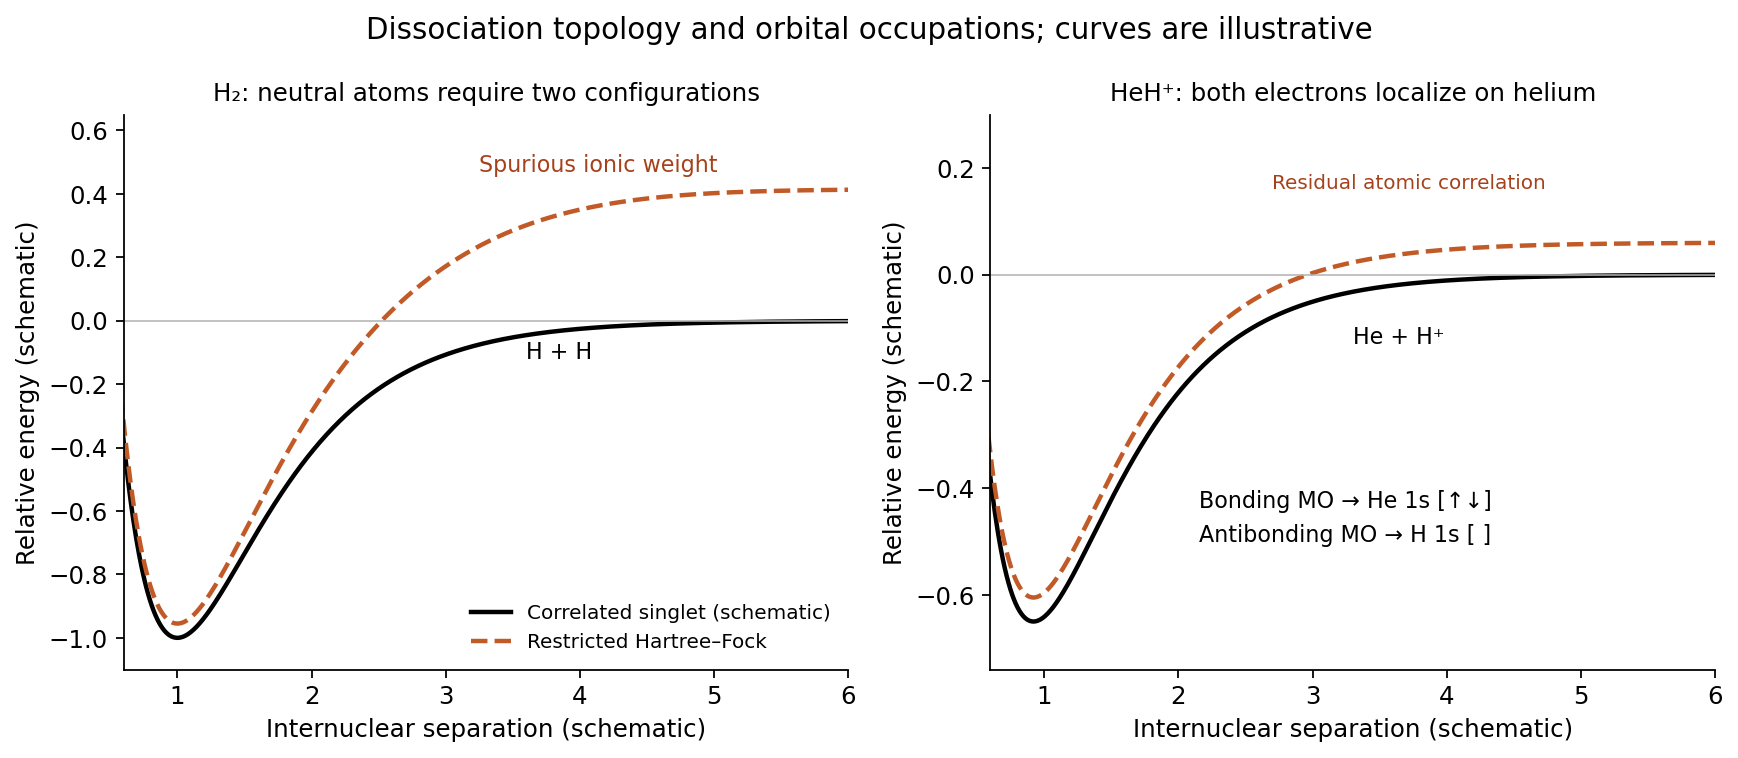


No: the lowest dissociation channel is , which a doubly occupied helium-centred orbital can describe. In the heteronuclear molecular orbital diagram the helium site lies below the hydrogen site. As separation grows, the bonding orbital tends to helium , the antibonding orbital tends to hydrogen , and the two ground-state electrons remain in the lower orbital.
There is therefore no symmetry requirement to keep both sites equally occupied and no compulsory half-weight mixture of wrong ionic configurations at dissociation. The physical asymptote is already a closed-shell atom plus a bare proton. The alternative limit lies higher. A Restricted Hartree-Fock method curve approaches the Hartree–Fock helium energy rather than the exact correlated helium energy, so ordinary dynamic electron correlation is still missing, but this is not the homonuclear static-correlation catastrophe. In the preceding two-panel sketch, the helium hydride curves have the same qualitative dissociation channel with a small residual correlation offset, whereas the hydrogen restricted curve reaches a qualitatively wrong high limit.
Articles by others on the same topic
There are currently no matching articles.