Polymer physics connects the conformations and motion of long molecules to the properties of a material. A polymer can change shape through many segment rotations while its chemical backbone remains intact. This makes the competition between flow and relaxation important: the same chain can behave as a coil in one experiment and an extended object in another. Soft matter includes both these flexible structures and liquid crystals, whose orientational order produces a different route to strong mechanical and optical response.
An idealized arrangement of confined DNA as approximately concentric loops around a central hole, with neighboring strands locally close-packed. It separates curvature-dependent DNA bending energy from interstrand interactions. It is a geometrical model, not a universal experimentally established arrangement in every capsid.
The change of an isolated flexible polymer from an expanded coil toward a dense polymer globule as its solvent becomes less favorable. In the Flory approximation, , where the positive three-segment coefficient prevents collapse to zero size. Positive gives swollen-coil scaling, gives the theta temperature condition, and negative favors a dense state with . Finite chain length rounds the change; polymer aggregation is a distinct multi-chain process and must be controlled in an experiment.
An interface changes polymer conformations through segment-surface affinity, solvent selectivity and confinement. Adsorbed chains make bound trains, intervening loops and free tails; end-grafted chains cross from isolated mushrooms to overlapping brushes as coverage rises. Surface binding energy competes with chain conformational entropy and crowding.
A polymer conformation model assigning allowed torsional states and statistical weights to successive backbone bonds. Correlations between torsional states can represent local stiffness beyond the fixed-angle, uniformly rotating freely rotating chain.
The effective volume governing a dissolved macromolecule’s hydrodynamic behavior and pore accessibility.
A dilute-solution bead-spring model including solvent-mediated hydrodynamic interactions. In its scaling regime, a chain of size has translational diffusion coefficient of order and longest relaxation time of order . Intermediate-time segment mean-square displacements scale as . Screening of hydrodynamic interactions in a concentrated solution changes the regime toward Rouse model behaviour; constraints from polymer entanglement require additional physics.
An overdamped bead-spring model of polymer dynamics with local friction and no long-range hydrodynamic interaction. For an ideal unentangled chain of identical frictional units, the centre-of-mass diffusion coefficient is proportional to and the longest internal relaxation time to . A tagged segment has an intermediate-time mean-square displacement proportional to before ordinary whole-chain diffusion dominates. Entanglement adds constraints beyond this model.
With additive drag from identical monomers, and centre-of-mass diffusion is . For monomer Stokes law , this gives . The model neglects long-range hydrodynamic coupling and does not alone describe the long-time spatial diffusion of a strongly entangled chain.
A polymer chain tethered by an end to a surface. Its surface number density sets a typical lateral separation . Comparing with the isolated coil size determines whether grafted chains behave almost independently or must stretch cooperatively away from the surface.
Formation of a tethered polymer layer by growing chains from reactive sites on a surface. Small monomers reach the growth sites without inserting an entire preformed coil, making high grafting density more accessible than in grafting-to. Initiator availability, termination and control of chain-length distribution remain important.
Formation of a tethered polymer layer by attaching reactive ends of preformed chains to a surface. The chains can be characterized before attachment, but inserting further coils into an existing polymer brush costs conformational entropy and produces a coverage-dependent barrier. Reaction chemistry and surface accessibility also limit the final grafting density.
The number of grafted chains per unit substrate area. For roughly uniform attachment spacing , it scales as .
A layer of sufficiently crowded tethered polymer chains stretched away from their attachment surface. In a good solvent, overlap increases the interaction contribution to free energy; extension relieves this crowding at a cost of chain entropy. The mushroom-to-brush crossover occurs at , where is an isolated coil size. In a simple good-solvent scaling model, the brush height is for effective segment excluded volume and statistical segment length ; if , this becomes . The scaling requires strong stretching relative to the isolated coil but extension well below the contour length .
A dense uniform polymer brush with must have height by polymer-brush volume fraction. The contour bound therefore requires . If , the limit is ; beyond it the putative incompressible branch demands chains longer than their polymer contour length. This is a failure of that microscopic packing model. A Kuhn length does not alone determine .
Conservation of material in a uniform polymer brush gives its mean volume fraction as , with material volume per counted segment, chain length , grafting density and height . Identifying with is an extra coarse-graining assumption. The Alexander-de Gennes brush model then gives , including powers in a good solvent and in a theta solvent.
In the Alexander-de Gennes brush model, a theta solvent gives . Stacking blobs then gives the displayed height, with mean polymer volume fraction for segment volume . Cancellation of the two-segment excluded volume does not remove three-segment repulsion or make the crowded layer freely interpenetrable at no cost.
A scaling model of overlapping end-grafted polymers assigns each chain a column of lateral width at grafting density . Each concentration blob contains statistical segments, and stacking blobs gives polymer brush height . For assumed segment volume , the mean volume fraction is . It assumes a long many-segment-blob brush and supplies scaling powers rather than a precise chain-end distribution.
End-grafted chains are approximately isolated when their lateral spacing exceeds their coil size. When the spacing becomes smaller, excluded-volume crowding favors extension normal to the substrate. The crossover criterion is a scaling estimate, not a sharp phase transition.
In a laterally uniform dense polymer brush with approximately fixed polymer volume fraction , conservation of occupied volume gives , with repeat-unit volume . At fixed solvent quality this gives , unlike the good-solvent brush height. Isolated collapsed polymer globules and laterally segregated grafted layers need not obey this uniform-layer model.
For an overlapping layer of long flexible grafted polymers, concentration blobs of width contain statistical units under the three-dimensional Flory approximation. A chain of blobs has height , giving the displayed good solvent scaling. Numerical prefactors are not supplied by this argument; extension must remain below the polymer contour length.
A dilute layer of end-tethered polymer coils whose lateral separation greatly exceeds the isolated coil size. Each coil retains approximately its isolated-solvent conformation, modified by the impenetrable tethering surface. Increasing the grafting density drives overlap and the crossover to a polymer brush.
A polymer entanglement is a constraint on a chain's motion caused by the inability of neighbouring chain contours to pass through one another. It is not a permanent chemical cross-link. In a dense polymer melt, these constraints can create long relaxation times and a transient rubbery plateau. Chains eventually release constraints through mechanisms such as reptation and motion of their neighbours. Entanglement therefore strongly affects dynamics even when the equilibrium size of a long chain remains approximately that of an ideal chain because excluded-volume interactions are screened.
A polymer melt is a dense fluid of polymer chains with little or no solvent. For a long flexible chain in a homogeneous melt, other chains screen its long-range excluded volume interactions: distant segments of the chosen chain displace statistically similar segments belonging to its neighbours. On scales above local stiffness, equilibrium chain dimensions are consequently approximately those of an ideal chain, even though monomers still cannot physically overlap. Entanglements can strongly affect dynamics without changing this leading large-scale equilibrium size law. Crystallinity, strong orientational order or unusual topological constraints require separate treatment.
The end-to-end vector joins the two ends of a polymer chain. In an isotropic unforced polymer solution, its vector expected value is zero; a useful measure of chain size is its root-mean-square length. For an ideal chain with independent isotropic steps of length , the cross terms in the square of the step sum have zero expected value, so . For a long Gaussian chain, its radius of gyration obeys . If counts chemical repeat units instead of independent Kuhn length segments, the statistical length used in must be calibrated to that choice of ; it is not automatically the physical bond length or the Kuhn length itself.
The path length along a polymer backbone. For links of length it is , whereas the end-to-end distance of a polymer chain cannot exceed it. A statistical segment count need not equal the chemical degree of polymerization.
The end-to-end distance of a polymer chain with fixed-length links of length cannot exceed its polymer contour length . A Gaussian chain spring has no such bound, so it eventually becomes an unphysical extrapolation. For a three-dimensional freely jointed chain under force , the mean extension along the force is , where . Its small- limit is , and its large- limit approaches from below.
Reptation is the snake-like motion of a long polymer through the constraints imposed by neighboring chains or a fixed porous matrix. A tube model restricts large transverse motion while allowing curvilinear motion along the chain contour. For a flexible chain with additive segment friction, contour diffusion scales as and the time to renew a contour of length proportional to scales as . With coil size squared proportional to , this gives a centre-of-mass diffusion coefficient proportional to . Applying this tube picture to weak-field DNA gel electrophoresis gives an inverse-length mobility over its appropriate regime; field-induced orientation and trapping can invalidate that simple law.
A tensile blob is the scale below which thermal conformations of a stretched polymer are weakly disturbed by tension . Balancing stretching work with gives . For free-chain size law , each blob contains monomers. A stretched string of blobs has extension and stretching free energy of order . This description requires blobs larger than individual statistical segments and smaller than the unperturbed coil.
For a long flexible polymer in a good solvent, a tensile blob of size contains statistical segments. Aligning blobs gives . The three-dimensional Flory approximation gives the displayed power. The regime requires : below its force range the coil has ordinary linear response, and above it finite link extension matters. Numerical amplitudes depend on solvent quality and segment conventions.[ref]
Excluded Volume Effects and Stretched Polymer Chains by Philip Alan Pincus. abstract and section I, p. 386
Polymer adsorption binds a polymer near an attractive interface through contacts of some of its monomers. The binding energy competes with the loss of conformational entropy. A chain need not lie flat: noncontacting loops and tails can extend into the solvent. In a weak-adsorption blob model, the layer thickness exceeds the Kuhn length and local chain statistics remain approximately those of the free chain below that thickness.
A nonadsorbed terminal section extending from a surface-bound part of an adsorbed chain to a free chain end. It has one surface connection, whereas an adsorbed loop has two.
A nonadsorbed chain section whose two ends connect to surface-bound portions of the same adsorbed polymer. Loops extend into the solvent and contribute to the thickness, steric repulsion and conformational entropy of an adsorbed layer.
A contiguous sequence of segments making contacts with an adsorbing surface. The adsorption energy gained per contact competes with the loss of chain configurations. A chain can have several trains separated by loops rather than lying wholly flat.
Let an adsorbed Gaussian chain gain free energy per bound monomer. A pulled segment of Kuhn length steps spanning height has elastic free energy , giving . Minimizing over yields and . Its height derivative is the constant peeling force displayed above. The plateau lasts while and the Gaussian chain approximation remains valid. In the slab model , optimization gives and .
An adsorption blob is a subchain of size comparable to the adsorbed layer thickness . If the free-chain size law is , it contains monomers. Confinement costs of order per blob produce . A simple contact fraction gives . This model predicts and binding gain per monomer of order . It assumes a many-blob layer and a monomer-scale attraction range; it does not replace the full surface-critical theory of polymer adsorption.
A characteristic time for a chosen polymer deformation to relax after forcing stops. Its value depends on the measured quantity and the chain model; one polymer can have multiple relaxation modes.
A coil-stretch transition is a sharp change from a coiled to a substantially extended polymer conformation under sufficiently strong extensional flow. The driving extensional strain rate must compete with the chain's relaxation rate, and the available residence time matters. A long polymer relaxation time lets a weak but sustained flow produce a large effect. The Hookean dumbbell in planar extension below isolates this competition; its unbounded growth signals a model limitation, rather than a prediction of infinitely long molecules.
Pierre-Gilles de Gennes's 1991 lecture describes Andrew Keller and collaborators using four rollers to deform a dilute polymer solution. A well-chosen exit trajectory kept coils under sustained extension. Above a threshold flow rate the solution became birefringent, revealing a change in chain orientation and conformation. The experiment connected an optical readout to a mechanically driven molecular response.[ref]
Soft Matter by Pierre-Gilles de Gennes. Printed p. 9: four-roller experiment and coil-stretch interpretation; text inspected in the Columbia reproduction
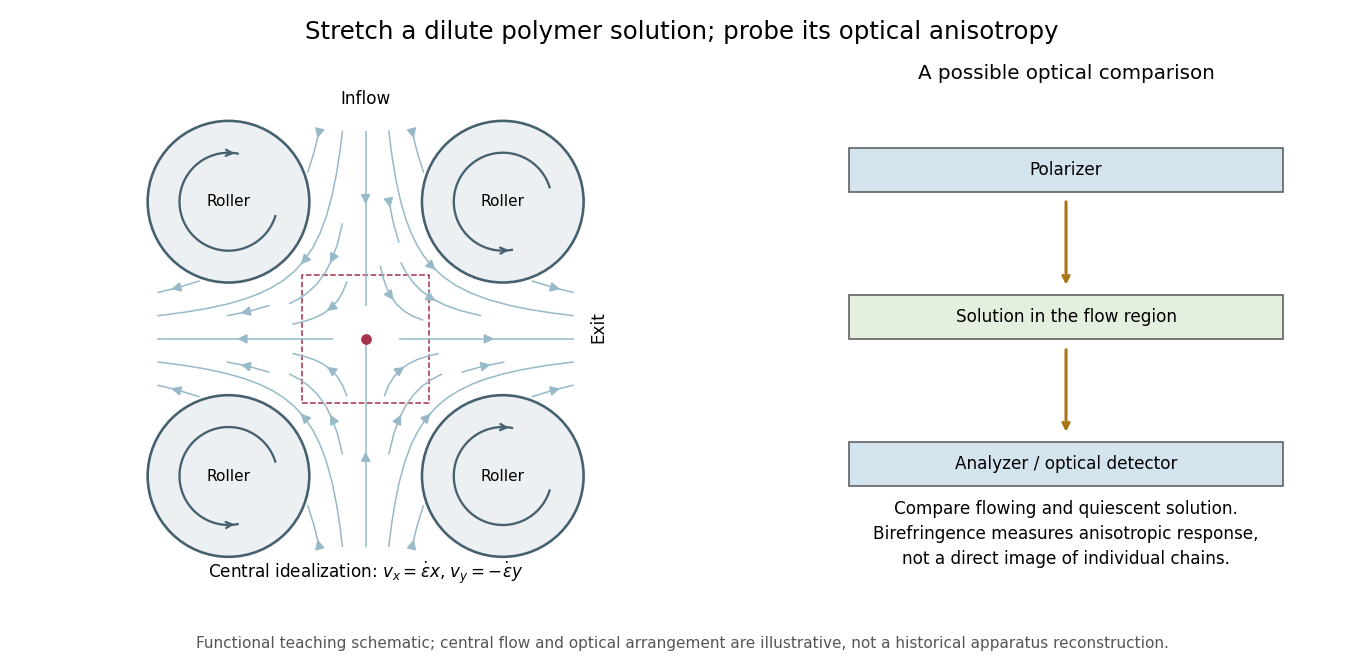
This original functional diagram shows a central flow idealization, rather than historical dimensions or a photograph. A probe of birefringence detects different optical responses along different directions; it does not directly photograph a chain or establish its extension without an optical model. Comparing a flowing solution with its quiescent state and controlling temperature and concentration are useful proposed checks, not an assertion that this is the complete historical protocol.
An original local flow model has velocity for constant extensional strain rate . Its divergence is zero, and its trajectories satisfy the displayed equations. Material approaches along and leaves along . If the idealized observation region ends at , a trajectory starting at has residence timeFor chosen , and , this is . Starting ten times farther from the symmetry line halves this particular residence time. The mathematical trajectory never exits in this model; diffusion, imperfect flow and finite apparatus prevent treating that ideal trajectory as a guaranteed infinite exposure in an experiment.
Represent one polymer by two beads joined by an ideal linear entropic spring, and stipulate a dimensionless conformation tensor with equilibrium value . The displayed model uses velocity gradient and positive polymer relaxation time . It omits finite chain extensibility, interactions with other chains, and feedback from the polymer onto the flow. It is a teaching model, not a fitted description of Keller's apparatus.
Define the Weissenberg number for this model as and dimensionless time . With an initially isotropic , the diagonal entries obeyBelow , both transverse and longitudinal responses approach positive finite steady values:For a chosen , and : the longitudinal mean-square separation is five times its quiescent value, while the transverse separation contracts. A root-mean-square length grows as the square root of a variance, so the longitudinal length factor is , not five. The anisotropy is an orientational indicator; converting it to measured birefringence requires an additional calibrated optical relation.
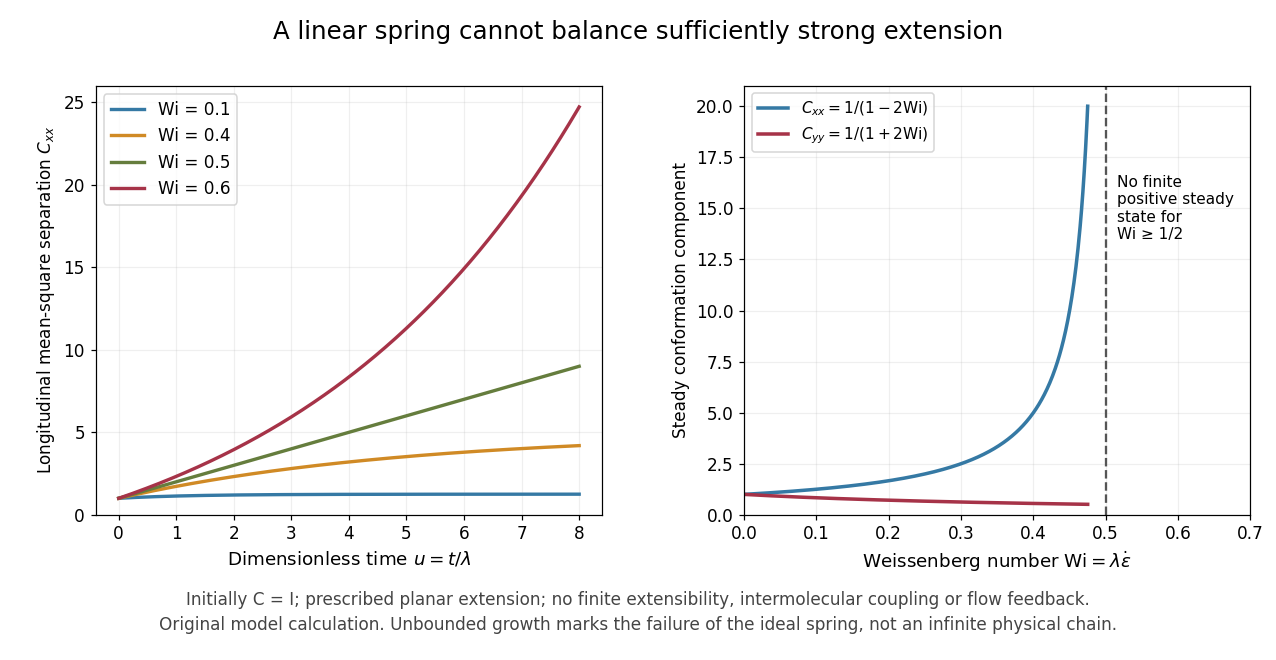
At , . Above that value, the growing homogeneous solution prevents a positive finite steady state. For instance, gives . Real chains have finite contour lengths, so this growth identifies the failure of the linear spring model. It does not establish a universal threshold for arbitrary flow histories, finite chains or other definitions of polymer relaxation time. Combining the growth equation with residence time in planar extension also explains why a large instantaneous Weissenberg number alone does not guarantee a strongly stretched exiting chain.
Articles by others on the same topic



Polymer physics is a branch of condensed matter physics that focuses on the physical properties and behavior of polymers—large molecules composed of repeating structural units known as monomers. Polymers can include natural substances like proteins and cellulose, as well as synthetic materials such as plastics and rubber. Key areas of study within polymer physics include: 1. **Structure and Morphology**: Understanding the arrangement of polymer chains and how their structure affects properties.