Useful proposed controls, rather than reported measurements, are a matched mixture omitting the initiating reagent, an unheated mixture, and a repeat of the complete formulation. Keep monomer amount, solvent, tube preparation and acquisition conditions matched where possible. The unheated comparison needs a readout that does not itself initiate polymerization; placing it in the hot probe would defeat that control. Amixture omitting iodine asks a different question: it changes chain transfer in chain polymerization and induction chemistry, so it is not necessarily a no-polymerization control. Record the pre-reaction baseline and compare crude material with isolated product; isolation can selectively remove shorter chains. These comparisons help separate initiation-dependent monomerconsumption from signal drift and unrelated chemical reactions.
The mechanistic inference is deliberately limited. Consumption plus independently characterized larger chain products supports polymerization; evidence of an induction stage helps testa proposed reaction mechanism. Neither readout directly counts stars in Pj∗, proves the absence of chain-growth termination, nor recovers every chain length. A chain-extension test would probe retained growth capability, but even a successful extension would not by itself establish that all original chains extended. No measured molecular distribution is reconstructed here. IUPAC cautions that reversible deactivation can produce living-like behaviour even when some termination occurs.[ref]
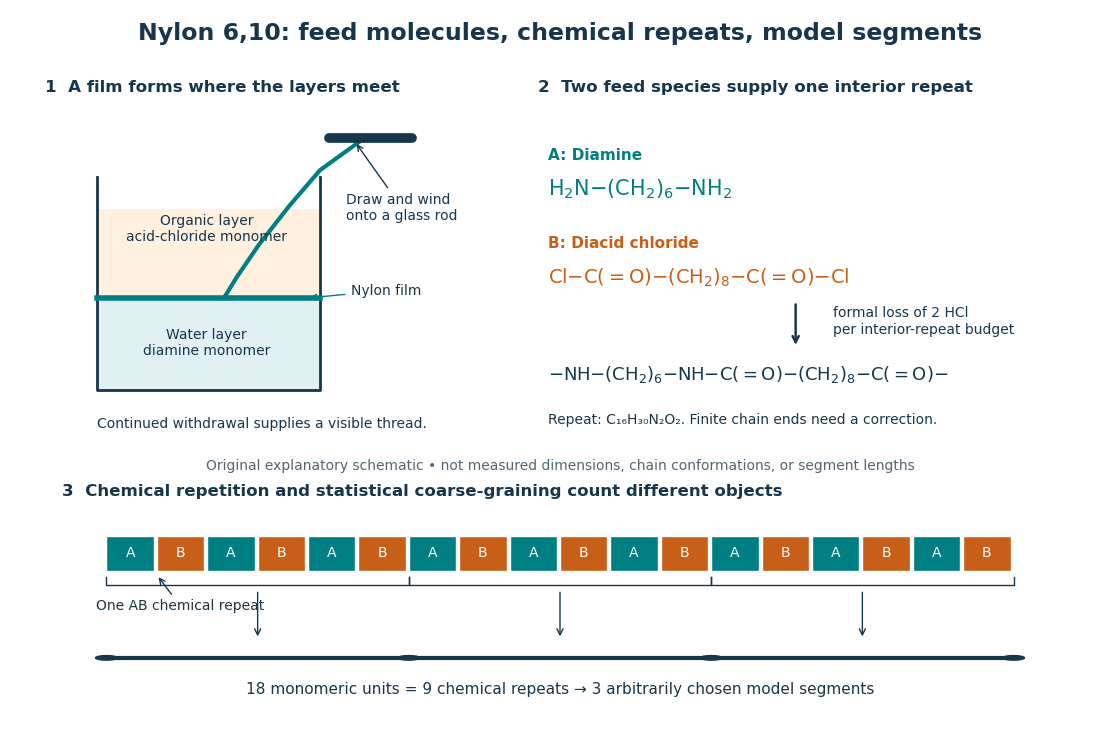
Its molecular formula is C16H30N2O2. The diamine feed is C6H16N2, and the diacid-chloride feed is C10H16Cl2O2; subtracting two HCl gives the interior-repeat budget. For a finite linear chain made from n of each reactant, there are 2n−1 joining connections, hence 2n−1 formal HCl eliminations, rather than 2n. One amine end and one acid-chloride end remain in that ideal unmodified chain. This is a stoichiometry check on the depicted reaction, not a claim that the washed experimental thread retains those ends. The reacting end-groupchemistry is documented in Hammond, lecture 5, p. 4; the specified Nylon 6,10reactantpair is reported in Polymer Day, nylon demonstration.
where the denominator includes the operative chain-transfer or chain-stopping rates per active chain. The Hf catalyst can propagate more slowly yet make longer chains if transfer is suppressed by a still larger factor. Slower insertion alone would not explain longer polymers. Distinguishing particular transfer pathways would require polymerend-groupanalysis and the concentration dependence of molecular mass and polymerization rate.
For a hollow microtubule, use I=π(ro4−ri4)/4 rather than the solid-cylinder value. For a clamped seed and amoment-free tip free to slide along the wall, linearized anglebalance is Bθ′′+Fθ=0, with θ(0)=0 and θ′(ℓ)=0. Its first nonzero solution has F/Bℓ=π/2, hence Fc=π2B/(4ℓ2). A different tip constraint changes this coefficient.
Increasing the unsupported length decreases that threshold. For growth against awall at fixed projected gap d, polymerization-driven cantilever postbuckling gives a more direct quantitative statement. The inextensible elastica obeys Bθ′′+Fsinθ=0. Multiplying by θ′ and using the zero tip moment gives
where the complete elliptic integrals are K(k)=∫0π/2(1−k2sin2ϕ)−1/2dϕ and E(k)=∫0π/2(1−k2sin2ϕ)1/2dϕ. Using K=(π/2)(1+k2/4+⋯) and E=(π/2)(1−k2/4+⋯) gives ℓ/d=1+k2+O(k4), and therefore
Thus the initial postbuckling force decreases as more contour is inserted. This result describes the weakly buckled branch before overhang, further wallcontact, or altered tip constraints invalidate the model. It is a mechanical forcelaw; a chemical force limit from polymerization can impose a smaller force.
For A, increasing concentration permits end-to-end intermolecular association to compete successfully with ring closure. It produces long, reversible supramolecular chains. Increasing chain length and entanglement account for the highly viscous solution without requiring covalent polymerization.
B has more favorably positioned attachment points on its rigid aromatic core. Its arms can close a less strained ring, so intramolecular association remains strongly competitive over the stated range. Predominantly discreterings do not form the same extended chain network, explaining the retained fluidity. Both compounds can make the same local hydrogen bonds; their different balance of ring strain, preorganization and intermolecular association explains the macroscopic difference.
. Original schematic of the authored UCSB polymerization demonstration described below. Chemical formulas identify the actual reactants and interior repeat; colored blocks identify their contributions. The lower coarse-graining sketch deliberately groups three repeats per model segment. Shapes, sizes, and that grouping are illustrative, not measured molecular geometry or a fitted Kuhn length. Details.
A motivating experiment: drawing nylon from aliquid interface. Feldwinn's UCSB demonstration places 25mL of an aqueous chemical solution of hexamethylenediamine, H2N(CH2)6NH2, in a 100mL beaker, then gently adds 25mL of sebacoyl chloride, ClC(=O)(CH2)8C(=O)Cl, in an organic solvent. The procedure specifies hexane; other parts of that handout list cyclohexane. The layers remain separate. The instructor catches material at their interface with tweezers and winds it slowly onto aglass rod; the handout reports continued spooling of Nylon 6,10. It specifies freshly prepared acid-chloride chemical solution and washing the thread before touching it. These are observations and conditions of a supervised laboratory demonstration, not measurements of chain lengths. See Feldwinn, UCSB Demonstration Library, Demo 044, pp. 1–3.
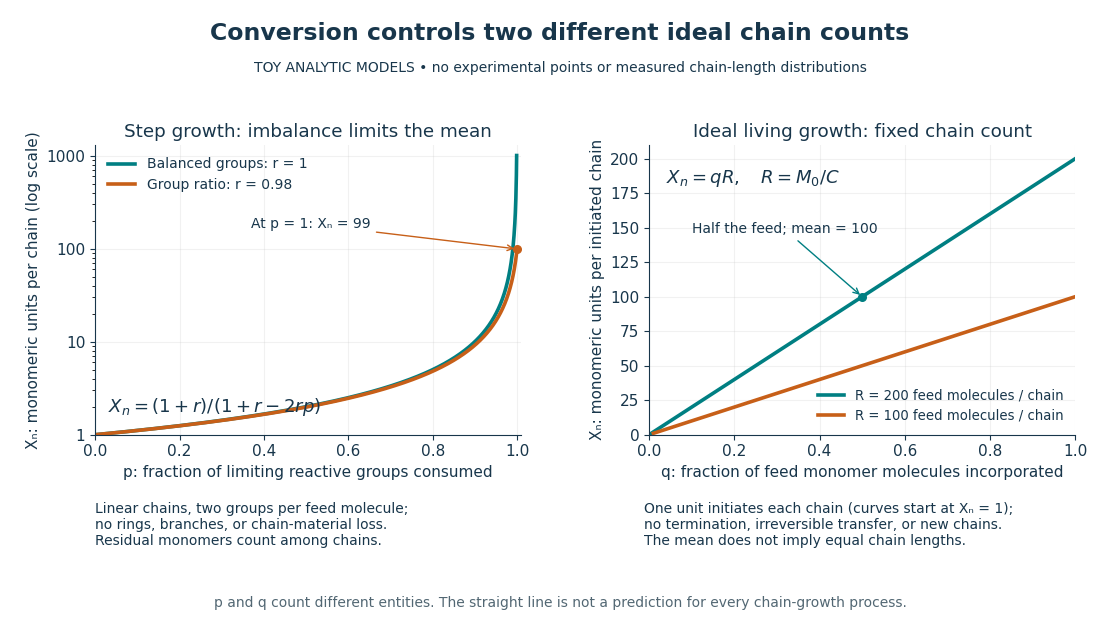
For an AA+BB feed with A0≤B0 initial reactive-group counts, put r=A0/B0 and now let p denote the fraction of the limiting Agroups consumed. Under the same linear, no-ring assumptions, N0=(A0+B0)/2 and N=N0−pA0, giving
. Toy analytic curves, not measured data. Left: Xn=(1+r)/(1+r−2rp), with p the limiting-group conversion and r the initial reactive-groupratio. Right: Xn=qR, with q the fraction of feed monomer molecules incorporated and R their initial number per initiated chain. The two horizontal axes count different entities; the right plot represents a special idealmodel, not all chain-growth polymerization. Details.
The useful connection is between local chemistry and a macroscopic material: selecting reactive ends and their proportions controls connectivity, while the resulting polymer's conformation is a separate problem described by models such as the freely jointed chain or Gaussian chain. Formation of chemical bonds, the count of monomeric units, and the count of statistical segments must be kept distinct.