A reaction mechanism describes how reactants become reaction products through specified elementary reactions. Adding the steps must recover the overall chemical equation; intermediate chemical species cancel. That accounting is necessary, but the proposed steps must also predict a rate law compatible with experiment. Several reaction mechanisms can pass the same test.[ref]
12.6 Reaction Mechanisms. Chemistry 2e by Paul Flowers and others; §12.6, opening discussion and Relating Reaction Mechanisms to Rate Laws: net equations, elementary steps, and experimental constraints
For example, the stipulated steps and sum to . I is a reaction intermediate: it is formed and then consumed. The catalyst C in the existing catalytic cycle instead starts available, participates, and is regenerated; its bound form can be a reaction intermediate. Both can disappear from the summed chemical equation, so cancellation alone does not distinguish their roles. Cancellation is algebraic bookkeeping, not an assertion that every step finishes instantaneously.
The net chemical equation supplies consumption ratios, not a clock. For the stipulated two-step example, directional reaction rates and give and . Initially can mean zero reaction product formation while A already disappears. In particular, requires negligible intermediate accumulation here. A single elementary reaction with the same net chemical equation would instead predict immediate formation at . Measuring both sides can reveal a distinction hidden by the net equation.
This makes reaction mechanisms useful for designing observations: follow a candidate reaction intermediate, measure individual reaction products, or vary one starting concentration at a time. The consecutive first-order reaction mechanism quantifies an intermediate's rise and fall; the parallel first-order reaction mechanism shows how identical reactant traces can conceal different reaction products. The acetone iodination rate-law experiment connects such reasoning to a real laboratory protocol. The existing reaction coordinate, potential energy surface, and energy profile describe configurations and barriers along a route; a concentration plotted against time describes a different observable.
A curved arrow whose tail marks an electron pair and whose head marks its destination in a proposed reaction step. The pair may start as a lone pair or a bond pair; a full arrowhead represents two electrons.
A consecutive first-order reaction mechanism passes material through a reaction intermediate. Consider a stipulated chemical reaction network , with constant positive rate constants, fixed temperature and volume, uniform mixing, and no reverse or competing chemical reactions. All three labels carry one conserved unit of material. Set and . The law of mass action givesAdding the three rate equations proves . Each step has a first-order reaction rate law; this does not make all three concentrations simple exponential decays.
For , the exact solution isTo derive the middle expression, substitute the first into the first-order linear differential equation for I and multiply by the integrating factor . Then , which can be integrated directly with the stated initial condition. The material balance supplies P without another integration.
The apparent division by zero at equal rate constants is removable. For , the same integrating factor givesThis is also the limit of the unequal-rate constant expression. Initially , so reaction product formation begins with zero slope despite immediate reactant loss. By contrast, a direct first-order reaction with the same has and nonzero initial slope. Their A traces agree exactly; their P traces do not.
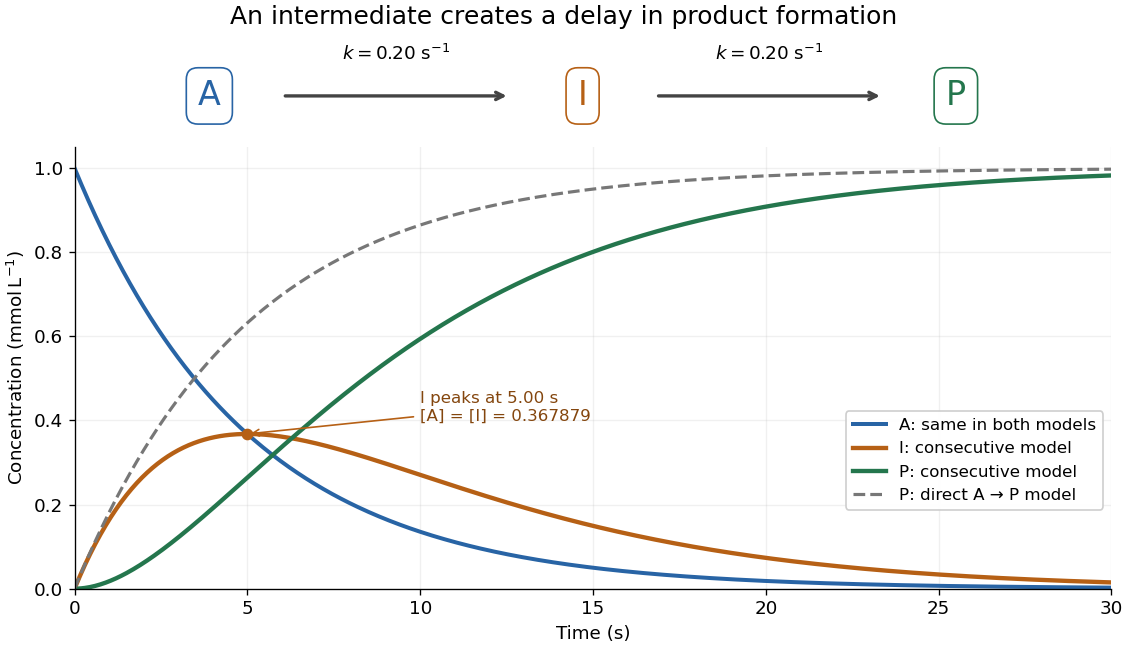
An accumulating reaction intermediate delays reaction product formation in a consecutive first-order reaction mechanism
. Original chosen model with and . Solid curves follow the exact three-species rate equations; the dashed P curve is a separate direct A-to-P model with the same A trace. These are model predictions, not measurements or an identified molecular reaction mechanism. Details.For the chosen equal-rate constant example, I peaks at , because . At that time and ; the direct model would already have . The peak means formation and consumption of I are momentarily equal, not that the whole system has reached chemical equilibrium.
More generally the peak is at , which is positive for either ordering of the two positive rate constants and tends to as they coincide. When , a brief initial adjustment can be followed by and approximately equal formation and consumption reaction rates. The exact solution states when this small-intermediate approximation can be reasonable. It is inappropriate for the illustrated equal-rate constant case: at the peak, over one third of the conserved material is I. Such timing matters if the goal is to collect a useful reaction intermediate before it becomes the final reaction product.
A parallel first-order reaction mechanism consumes the same reactant through competing routes. Extend the product-split idea in catalytic selectivity by following both reaction products over time. Stipulate irreversible first-order reactions and in a uniformly mixed closed system of fixed volume and temperature. Initially only A is present, with concentration . With ,Here abbreviates a sum of rate constants, not an equilibrium constant. The rate equations are , , and . Integrating the last two after solving the first proves the expressions and their conserved total .
Choose two alternative models: model U has ; model V has . Both have , so every measurement of A alone is identical, even with no measurement error. Yet their P-to-Q concentration ratios at any are and , respectively. This is a precise failure of identifiability from that one observable. A calibrated measurement distinguishing P from Q resolves it within the stipulated two-route model.
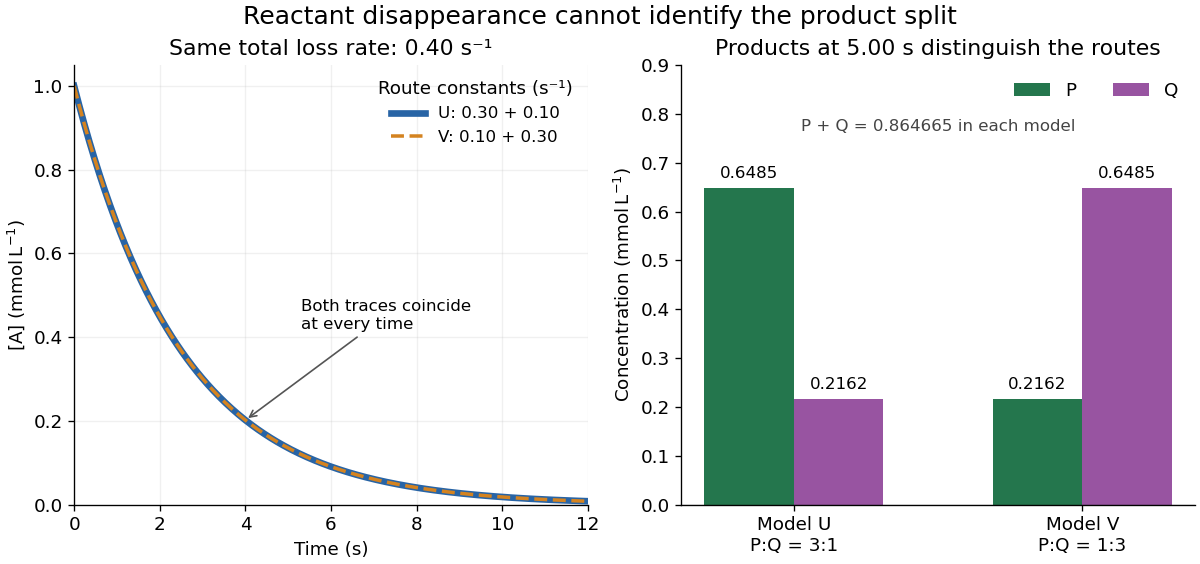
At , both models give . Model U gives ; model V exchanges these. Measuring only the summed reaction product concentration would also fail to distinguish them. Product-specific observations therefore answer a question that total conversion cannot. As with any reaction mechanism, fitting these observables still does not rule out a more elaborate untested route.
Acetone reacts with iodine to form iodoacetone, iodide, and a hydrogen ion:In water, denotes a solvated hydrogen ion; added hydrochloric acid provides acidic conditions.
Moorpark College's protocol mixes acetone, hydrochloric acid and water from burets in matched test tubes; pipet in iodine, swirl and time colour loss against white paper. Use water, then reacted solution, as references. Duplicate each mixture; keep total volume and run temperatures within .[ref]
Experiment 4: Chemical Kinetics: The Iodination of Acetone. Chemistry M01B Laboratory Manual, version 4.0 by Moorpark College Department of Chemistry; Version 4.0, Experiment 4, printed pp. 17–18: Procedure and Table 1
The stock dilutions give , , and . B and C separately double acetone and acid; D halves iodine. The manual supplies no measured times; it states zero partial order of reaction in iodine and asks students to determine the other exponents.[ref]
The endpoint measurement is a time to an operational colour threshold, not direct observation of a reaction intermediate. If the iodine disappearance rate is approximately constant until that endpoint, . Halving at fixed other concentrations then approximately halves without changing . A longer or shorter endpoint time alone is not a comparison of reaction rates unless the starting concentration is included. For an observed power-form rate law, doubling only acetone gives a rate ratio and hence ; doubling only acid similarly estimates its own partial order of reaction. Those are inference formulas, not asserted experimental results.
A useful proposed reaction mechanism would explain how iodine is consumed after an earlier transformation sets the throughput, making the observed reaction rate insensitive to iodine over a tested range. The observed zero exponent would contradict treating this net chemical equation as a single elementary reaction with directional rate law . It would not uniquely prove which reaction intermediate or earlier step is involved. Independently detecting a candidate reaction intermediate would provide another kind of evidence.
The excess-reactant approximation has an explicit scale: the baseline iodine inventory is only of the initial acetone inventory, and its net hydrogen ion production is of the initial acid inventory, assuming only the stated one-to-one chemical reaction. These are calculated bounds on composition change, not measured reaction orders. Mixing delay, a subjective colour threshold, and temperature drift limit the inference. Repeats and a fixed viewing geometry help; a calibrated absorbance trace could test linearity during conversion instead of relying on an endpoint alone. That extension would require a reference blank and a justified Beer-Lambert law calibration, rather than treating visible colour as an exact concentration. A zero-order reaction approximation in iodine must fail at depletion; it cannot imply consumption after no iodine remains.
A reaction intermediate is formed during a reaction mechanism and consumed in a subsequent step, cancelling from the net chemical equation. It can accumulate, as I does in the consecutive first-order reaction mechanism. A reaction intermediate can also connect transformations in energy coupling. Unlike a catalyst, it need not be present before conversion begins.[ref]
12.6 Reaction Mechanisms. Chemistry 2e by Paul Flowers and others; §12.6, opening ozone example: intermediate formation, consumption, and cancellation
A reaction intermediate is distinct from a transition state: the former can persist between steps, whereas the latter describes passage through a barrier. The existing potential energy surface and energy profile provide the configuration-space picture; the time-dependent concentration of an intermediate instead measures an ensemble of reacting entities. Low concentration or short lifetime can make detection difficult, so failure to detect a proposed reaction intermediate is not by itself proof of absence. Conversely, detecting a chemical species does not establish that it carries the proposed route to the final reaction products; it could belong to a competing route.
A tetrahedral intermediate has a formerly trigonal carbonyl group carbon bonded to four groups after nucleophilic addition. Its subsequent elimination can restore a carbonyl group.
A reaction coordinate parametrizes changes in molecular configuration along a chemical reaction path. It can involve a bond length, a bond angle, or a combination of geometric changes. It is not elapsed time and is not the bulk extent of reaction: one describes configurations, while the other accounts for how much material has converted. The precise choice matters when interpreting an energy profile. See the IUPAC Gold Book: reaction coordinate.
A hypersurface separating selected reactant and product regions. A rate derived from signed flux and a suitable long-time commitment is independent of its location when the dynamics and normalization are treated consistently.
A potential energy surface assigns potential energy to the configurations of the reacting chemical atoms, for a specified electronic description. A reaction coordinate selects a path through this many-dimensional configuration space; a potential-energy profile then records the potential energy along that path. Different paths can share endpoints but cross different barriers. See the IUPAC Gold Book: potential-energy reaction surface.
A correlation between many-electron states classified by total spin and spatial symmetry along a nuclear path. Orbital occupations, spin adaptation and configuration interaction are needed; one-electron energy curves alone are not a state diagram.
A degeneracy between adiabatic electronic states with characteristic splitting in suitable nuclear-coordinate directions. It can connect to Jahn-Teller dynamics in open-shell triangular molecules.
An energy profile plots an explicitly specified energy against a reaction coordinate. Identify whether the vertical axis is potential energy, enthalpy, or Gibbs free energy; these descriptions are related but not interchangeable. A potential-energy profile describes a path through configurations, while a free-energy description also accounts for the relevant thermodynamic conditions.
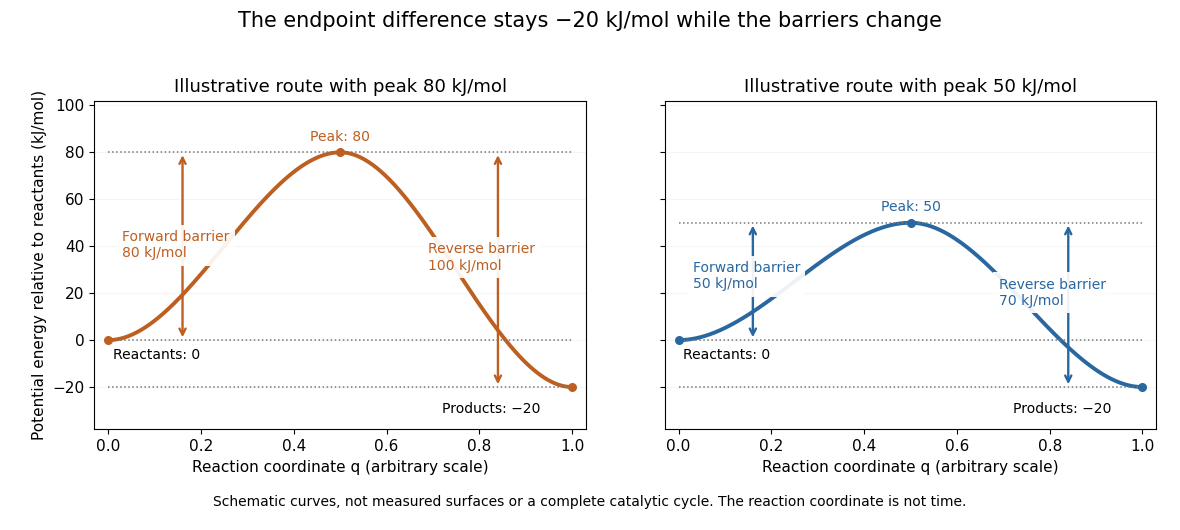
Two schematic potential-energy profiles share endpoints but have different barriers
. The reactant level is , the product level is , and the two maxima are and . These chosen curves illustrate endpoint differences and barrier heights; they are not measured surfaces, trajectories in time, or a complete catalytic cycle. Details.For the two chosen profiles, the reaction energy barriers from the reactants are and . From the reaction products, the same maxima are and higher. Both paths have endpoint difference . The barrier is measured from the relevant starting endpoint, not from an arbitrary zero of potential energy.
The appeal of the diagram is that it separates two useful questions: which endpoint is lower, and which path is easier to traverse? Catalysis changes the route through transition states while retaining the endpoint thermodynamics of the same overall chemical reaction. A real catalytic route may have several steps and reaction intermediates; the two simple curves do not specify such a reaction mechanism. The plotted potential-energy difference alone is not an equilibrium constant or a calculated rate constant. See Chemistry 2e, 12.7: alternative reaction routes and catalysis.
A plot of Gibbs free energy along a specified reaction coordinate under stated conditions. A barrier must be measured from the appropriate starting minimum.
A potential-energy profile plots the potential energy of the reacting system along a specified path from reactants through a transition state to reaction products. It is a particular kind of energy profile. For an elementary reaction, a suitable reaction coordinate parametrizes the path; a multi-step reaction mechanism needs the coordinates of its successive steps. See the IUPAC Gold Book: potential-energy profile.
A reaction energy barrier is the height of a specified peak relative to the starting state in an energy profile. State which energy is used and which direction is being considered. A schematic barrier is not automatically the empirical activation energy obtained from the temperature dependence of a rate constant.
An elementary reaction is one microscopic step in a reaction mechanism, with no intervening reaction intermediate needed within that step. Its molecularity counts the participating reacting entities. In an ideal dilute, uniformly mixed model, the law of mass action gives the directional reaction rate from the step's reactant concentrations: has , while has . Applying those powers to a net chemical equation requires separate evidence that it is elementary.[ref]
12.6 Reaction Mechanisms. Chemistry 2e by Paul Flowers and others; §12.6, Unimolecular Elementary Reactions and Bimolecular Elementary Reactions: step equations and directional rate laws
For a reversible step , apply that rule separately: and , giving net production of I as . Their difference can vanish at chemical equilibrium while both directional reaction rates remain nonzero. For , the elementary normalized reaction rate is , whereas A's disappearance rate is ; see second-order reaction for the complete factor-of-two example.
Calling a step slow does not remove the need for its actual reactant concentrations. If a reaction intermediate enters its rate law, determine that concentration from the other rate equations or a justified approximation. In the consecutive first-order reaction mechanism, an intermediate maximum is only one instant at which its production and consumption match. Replacing its entire evolution by that one balance would lose the initial buildup and delayed reaction product formation. A successful overall first-order reaction fit likewise does not establish an elementary event: both worked reaction mechanisms above provide ways for microscopic detail to remain hidden in one reactant trace.
Molecularity counts the reacting entities in one elementary reaction: one for a unimolecular step, two for a bimolecular step. An entity can be a molecule, ion, or chemical atom. Solvent entities involved only through solvation are excluded. Molecularity describes a microscopic event; order of reaction describes the concentration dependence of a rate law. Assigning molecularity to a net multi-step chemical equation is therefore inappropriate.[ref]
Glossary of Terms Used in Physical Organic Chemistry (IUPAC Recommendations 1994): M entries by International Union of Pure and Applied Chemistry. Glossary of Terms Used in Physical Organic Chemistry, IUPAC Recommendations 1994, entry “molecularity”: elementary event and exclusion of solvation medium
Articles by others on the same topic



A reaction mechanism is a detailed description of the steps involved in a chemical reaction. It outlines how reactants transform into products at the molecular level, including the sequence of elementary reactions, the formation of intermediate species, and the transition states that are formed during the process. Understanding the reaction mechanism helps chemists predict the outcome of reactions, optimize conditions for desired products, and design new reactions.