The incident X-ray drives the electrons of an atom. Away from an absorption edge, and when the photon energy greatly exceeds the relevant electronic binding energies but is small compared with , each electron has approximately the same Thomson scattering amplitude. The much heavier nucleus makes a negligible contribution because its acceleration is much smaller.
For elastic scattering, the amplitudes from different bound electrons add coherently. In units of the single-electron amplitude, the atomic scattering factor is the Fourier transform of the bound-electron number density:Consequently, when the electron cloud is not well resolved by the scattering vector, its internal phase differences are small andThis is an approximation to the coherent scattering amplitude, not an assertion that independently scattered intensities add as . At larger momentum transfer, destructive interference within the cloud reduces ; near absorption edges, the atomic scattering factor acquires dispersive corrections. Inelastic scattering is a separate contribution. In an ionic crystal, use the ion's bound-electron count rather than automatically using the neutral atom's atomic number.
Use the total scattering angle . For a cubic unit cell, Bragg's law and the Miller indices giveThe four measured values of , normalized to the first, are . Multiplying by three gives approximately , the initial allowed sequence for a face-centered cubic lattice:The FCC lattice factor is ; it vanishes unless all three indices have the same parity. Thus the initial ratios differ from those of a primitive or body-centered cubic lattice. This is the conventional minimal cubic indexing; arbitrarily hiding additional weak reflections could make a short list alone ambiguous.
The low-pressure rock salt structure has an FCC Bravais lattice with one NaCl formula unit per primitive unit cell, or four per conventional cube. A possible atomic basis of a crystal is Na at zero and Cl at in conventional fractional coordinates. The two species have unequal atomic scattering factors, so both the odd and even FCC families can occur.
The four independent lattice constant estimates areTheir small scatter reflects the rounded angular data. The appropriate reported result is
The high-pressure atomic basis of a crystal has the caesium chloride structure. Its chemically labelled Bravais lattice is simple cubic lattice: exchanging Na and Cl is not a translation symmetry. Its structure factor isBecause ionic bonding gives approximately Na and Cl, their bound-electron counts are and . In the stipulated electron-count approximation, odd-index-sum reflections have , while even-index-sum reflections have . In particular, the first ring is weak but present; it must not be discarded using a body-centering extinction rule.
The next ring is , and Bragg's law givesThe powder diffraction multiplicities are six and twelve. Counting all symmetry-equivalent reflections, while neglecting other angular factors, therefore givesThe corresponding single-directed-reflection ratio would be . Neutral-atom counts would give the wrong contrast because the material is ionic.
For measured integrated photon counts, the geometry must also be specified. More generally the ratio is times the ratio of Lorentz-polarization factors, thermal-motion factors and detection efficiencies. For example, the common unpolarized symmetric-powder convention hasand gives if all other corrections are suppressed. Thus 24.5 is the multiplicity-and-structure-factor estimate, rather than a geometry-independent precision count ratio; actual angle-dependent ionic form factors are also not supplied.
The high-pressure lattice constant follows from its first reflection:Conserve the number of NaCl formula units. The low-pressure conventional unit cell contains four, whereas the high-pressure cell contains one. The volume per formula unit changes from to , soThe sample contracts by about . Comparing directly with would compare different amounts of material.
The Debye model treats a three-dimensional solid as an isotropic elastic continuum with three independent acoustic phonon polarizations. It assumes harmonic normal modes, a linear dispersion relation , and equilibrium Bose-Einstein statistics with zero phonon chemical potential. Real anisotropy and nonlinear dispersion are replaced by one effective Debye sound speed. A cutoff restores the finite number of atomic vibrational degrees of freedom; the continuum itself would otherwise have infinitely many modes.
For sample volume , the wavevector state density is per polarization. Counting a sphere givesThere are modes for atoms. HenceThe associated phonon density of states isIf longitudinal and transverse sound speeds differ, use for the low-frequency count. The Debye frequency is an effective mode-counting cutoff, not a claim that every actual branch terminates at exactly that frequency.
The FCC conventional unit cell has four atoms, so the cube containsatoms, neglecting surface corrections. A phonon is one excitation quantum of a vibrational normal mode; there can be many quanta per mode, and zero-point motion does not add half a phonon to the thermal population.
For the low-temperature thermal phonon number, integrate the Bose-Einstein distribution against the Debye model phonon density of states:At , , so extending the upper limit to infinity has negligible effect at the precision required. The supplied integral yieldsThis is the thermally excited phonon number, not the number of available modes.
The continuum approximation fails when the thermally occupied wavelengths become comparable with the cube size . From the Debye temperature,A lowest nontrivial acoustic frequency is of order ; a standing-wave estimate corresponds to . Periodic boundary conditions instead give and . The precise spectrum depends on mechanical boundary conditions and branch speeds, neither of which is specified.
This finite-crystal phonon freeze-out requires summing discrete normal modes. For a finite positive lowest frequency, , rather than extrapolating the bulk law indefinitely. Rigid translations and rotations of an isolated crystallite are not internal thermal phonons. Surface acoustic modes can modify the intermediate crossover, but do not justify inventing a unique boundary spectrum.
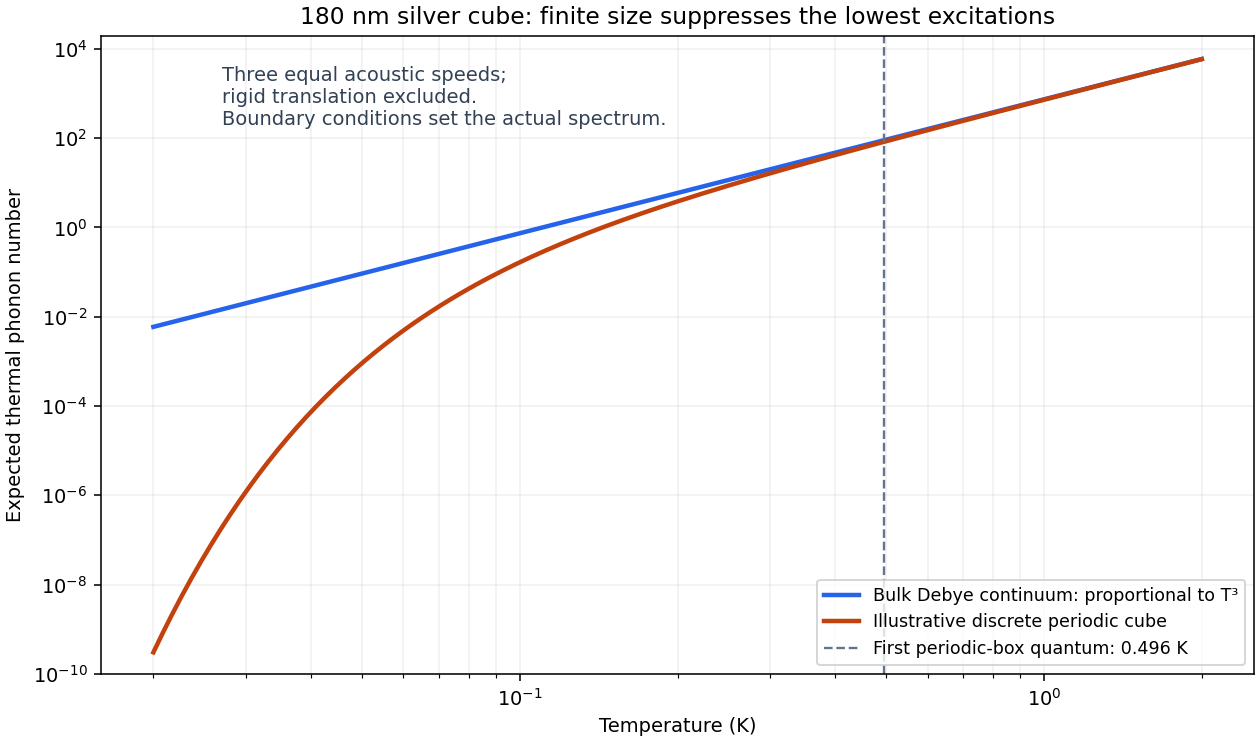
Bulk and finite-box thermal phonon populations
. The discrete curve uses an illustrative periodic cube and three equal acoustic speeds; its low-temperature suppression is not a fitted free-surface spectrum. Details.
The harmonic phonon contribution to the internal energy isFor , every retained mode has . Expanding cancels the explicit half quantum and gives per mode to leading order. ThereforeEquivalently, differentiating before integrating givesThe integrand tends to , and its integral is , producing the same Dulong-Petit law. In classical language, the equipartition theorem assigns each to the kinetic and elastic parts of every harmonic normal mode. Fixed zero-point energy does not contribute to the heat capacity.
Assume thermal equilibrium, nondegenerate carriers, and isotropic parabolic band dispersions, with the supplied effective masses interpreted as density-of-states masses. Take one effective valley with two electron spin states for each band; any additional valley multiplicity must instead be absorbed into the masses or explicit degeneracy factors.
Let the conduction band edge be , the valence band edge , and . The spin-summed conduction density of states per volume isUsing the nondegenerate limit of the Fermi-Dirac distribution and the substitution givesThe factor in the supplied integral produces the normalization. Counting missing valence band electrons similarly givesMultiplication cancels the chemical potential, establishing the semiconductor mass-action law:The result does not require an intrinsic semiconductor: semiconductor doping changes and separately while preserving this product under the stated equilibrium and nondegeneracy assumptions.
A substitutional group-V atom such as phosphorus has one more valence electron than germanium and acts as a donor impurity, giving an n-type semiconductor. Approximate its outer electron by a hydrogenic donor state with screened Coulomb attraction and effective mass . The ionization energy is the positive binding-energy magnitudeIts effective Bohr radius of a hydrogenic donor is about , much larger than an atomic core, consistent with a shallow extended bound state.
At , , while the mean three-dimensional classical translational kinetic energy is . Both exceed the donor binding energy, so substantial thermal ionization is expected, unlike excitation across the much larger host band gap. A precise ionized fraction also depends on the chemical potential, dopant density and donor charge-state degeneracy; comparing two energy scales alone does not prove complete ionization at arbitrary concentration.
With the supplied fixed band gap and effective masses, the intrinsic carrier concentration isNumerically,For uncompensated, approximately ionized donor impurities, charge neutrality and the semiconductor mass-action law requireThere is no abrupt extrinsic-to-intrinsic transition. An order-of-magnitude lower crossover density is thereforeRobust extrinsic dominance requires ; at the nominal equality, and , so intrinsic contributions are already appreciable. For a quantitative tolerance , the minimum ionized dopant density is . For example, would require at the upper temperature.
Using the nominal crossover estimate and the complete-ionization approximation at room temperature, andFor the twofold neutral-donor degeneracy, a consistency calculation gives about ionization at , raising this minority density to , about above the complete-ionization estimate. Stronger extrinsic dominance uses more donors and correspondingly fewer electron holes. Actual germanium has temperature-dependent band parameters and multiple valleys; the numerical estimate consistently uses the fixed effective data supplied rather than adding an unsupplied multiplicity.
A Wigner-Seitz cell is the region of real space at least as close to a chosen Bravais lattice point as to any other. It is bounded by perpendicular bisectors of vectors to neighboring lattice points and is a primitive unit cell.
A reciprocal lattice vector satisfies for every lattice translation . With primitive vectors , reciprocal primitive vectors satisfy , and for integers .
The first Brillouin zone is the Wigner-Seitz cell of that reciprocal lattice, centered at . Its boundaries are Bragg planes, . Higher Brillouin zones are the regions separated from the origin by successively more such planes: an interior point of zone has crossed exactly planes. A second Brillouin zone is generally disconnected in this extended-zone representation; it is not just a translated first-zone rectangle.
For area , allowed wavevectors have density . Include two electron spin states and fill a disk of radius at zero temperature. ThenThe free electron model has parabolic dispersion , with its minimum chosen as zero, soThese are the Fermi wavevector and Fermi energy for a spin-degenerate two-dimensional free electron gas. An imposed spin polarization or additional valley degeneracy would change the state count.
For the first two Brillouin zones of a rectangular lattice, write and , so and the reciprocal lattice vectors are . The perpendicular bisectors to the four nearest axial reciprocal points encloseThe second Brillouin zone consists of points beyond exactly one Bragg plane. In the first quadrant, it is the union of the following two regions:Reflect these regions in both axes. The first is a trapezoidal side lobe; the second is half of a triangular top lobe. The sloping boundary is the bisector for . On the side lobe, is the bisector condition for : crossing that plane too would enter the third or a higher zone. These additional bisectors are why four rectangles pasted onto the first zone would be incorrect.
Equivalently, the full right lobe has verticesand the top lobe has vertices . Add their left and bottom reflections. Their total area isequal to the first-zone area, as required by reciprocal-space state counting. These formulas remain valid for every , including very elongated rectangles; boundaries themselves can be shared by convention.
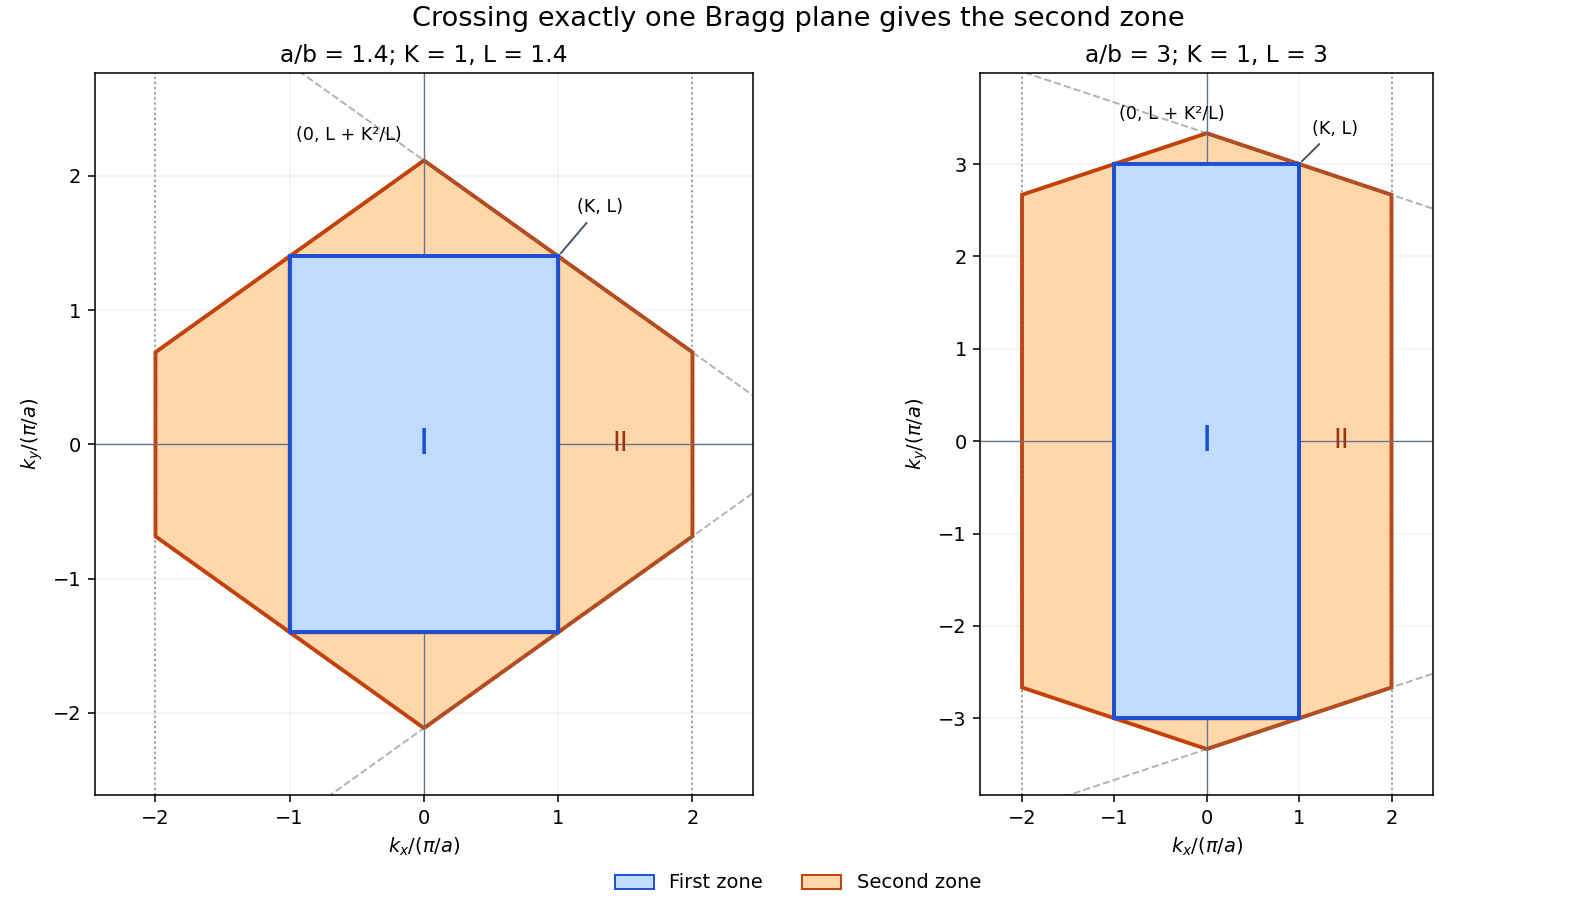

For one electron per primitive unit cell, the areal number density is . The Fermi circle has radius . The nearer first-zone boundary is , because . The circle protrudes precisely whenAt equality the Fermi surface only touches the boundary. Crossing an extended-zone boundary alone does not make a gap: the free electron gas remains metallic when the ionic periodic potential is neglected.
With spin degeneracy, each energy band contains two electrons per primitive unit cell. Divalency therefore permits a filled first band, but only if its maximum lies below the second band's minimum. With negligible ionic periodic potential, the circular Fermi surface and the rectangular zone have equal enclosed area, not equal shape. Electrons occupy some states outside the first zone while some higher-energy first-zone states remain empty. Folded bands overlap, and the material is a metal.
A periodic potential mixes plane waves differing by a reciprocal lattice vector. At a two-wave Bragg point, the nearly-free electron model gives a direct gap . Increasing appropriate Fourier coefficients can remove the band overlap. With the first band filled, the system is then a band insulator, provided the temperature is small compared with the insulating band gap in energy units.
The rectangular free-electron band-overlap scale follows from two different points in reciprocal space. Put . Before gap opening, the highest first-zone energy is at the corner, while the lowest second-zone energy is at the midpoint of the nearer face:Thus the downward shift of the first-band maximum plus the upward shift of the second-band minimum must overcomeIn a rigid-splitting estimate, an opening lowers the first band by and raises the second by , giving the familiar threshold . Equality is only band touching, not a positive insulating gap. This expression is the overlap that must be removed, under that explicit band-shift approximation.
Positive direct gaps can coexist with band overlap: a local direct splitting and the global indirect band gap are not interchangeable. The shape and Fourier coefficients of the ionic potential are unspecified, so there is no universal exact critical direct-gap width determined by alone. For a concrete demonstration, a separable potential has axial direct gaps and . To first order, the corner's lowest level is , and the two candidate second-band minima are and . Therefore, within this model and neglecting higher-order band shifts, nonoverlap requiresEqual axial gaps would give the first-order estimate , rather than the rigid-splitting estimate. Such a large opening can exceed the quantitative range of weak-potential perturbation theory, but the example identifies the missing model assumption: corner levels feel both couplings. The exact general answer is the positive indirect-gap criterion above; a geometry-only direct-gap threshold would claim more than the supplied potential information supports.
Articles by others on the same topic
There are currently no matching articles.