The chemical potential measures how a thermodynamic potential changes when matter is added under specified constraints. It plays for matter exchange the role that temperature plays for heat exchange: a difference supplies a direction, and equality characterizes thermodynamic equilibrium when the corresponding exchange is allowed. This connects diffusion, phase coexistence and chemical equilibrium through one quantity.
For a single component, the particle convention iswhere is internal energy, counts particles, is entropy and is volume. This partial derivative holds entropy and volume fixed; it measures an incremental change rather than the average energy per particle. See David Tong, Statistical Physics, section 1.4.
In chemistry the molar convention instead uses amount of substance , measured in moles:Here is Gibbs free energy, is temperature, is pressure and is the Avogadro constant. The units are per particle versus . For several chemical species, define each chemical potential by varying only its own particle count or amount of substance, with all other components' counts or amounts fixed. Many texts use for either convention, so the units and the variable being differentiated must identify it.
For a simple compressible system with variable component inventories, the fundamental differentials areThey describe changes between states in thermodynamic equilibrium with only pressure-volume work. If every component inventory is fixed, the sums vanish. The Gibbs free energy form follows by differentiating and using ; this is why the two definitions describe the same chemical potential in different units.
A chemical potential can be negative. Holding entropy fixed while adding matter need not increase internal energy; Tong explains this explicitly for a classical ideal gas in Statistical Physics, section 2.2.4. Its absolute sign also depends on the reference energy: replacing by shifts by . A common reference shift cancels from differences for the same chemical species, which are what determine exchange. Thus “energy cost” means a signed increment, not an always positive amount that must be supplied.
Water activity measures the chemical potential of water relative to a chosen pure-water standard state. Under ideal-vapour conditions above a flat solution, it is the equilibrium vapour pressure divided by that above pure liquid water at the same temperature. Dissolved solute usually lowers activity. Raoult's law gives the ideal dilute-solution limit; nonideal interactions require an activity model.
The chemical potential assigned to a specified standard state. With dimensionless thermodynamic activity , the per-particle convention is ; a molar convention uses . The standard state changes both and the activity definition without changing observable equilibrium.
For one internal species, the grand canonical partition function of a classical ideal gas satisfies , with quantum gas fugacity and thermal de Broglie wavelength . Differentiating the grand potential gives , hence . An internal partition factor must be included consistently if internal states are available.
A quasi-Fermi level is a local electron or hole population parameter in a semiconductor driven out of equilibrium, assuming each population is internally close enough to thermal equilibrium to use a Fermi-Dirac distribution. Separate electron and hole parameters replace one common equilibrium chemical potential. Their spatial profiles depend on transport, generation and recombination as well as electrostatics. For ideal ohmic contacts with the p voltage relative to n, the contact electron electrochemical potentials obey .
The difference between electron and hole quasi-Fermi levels under nonequilibrium conditions. For ideal selective contacts their terminal separation sets ; poor contacts can lose part of the internal splitting.
Consider two internally equilibrated reservoirs A and B at the same imposed temperature and pressure. They may have different compositions or represent different phases. Transfer a small positive amount of substance of the same chemical species from A to B, holding the other inventories fixed. To first order, their total Gibbs free energy changes byHence exchange toward lower chemical potential lowers Gibbs free energy. If transfer is allowed in both directions and the two reservoirs contain the exchangeable component, thermodynamic equilibrium requires equal chemical potentials for that component. See MIT 5.60, Lecture 14, pages 1–2.
An impermeable partition can maintain unequal chemical potentials; equality is a condition on an allowed exchange. Two pure samples of the same ideal gas already have equal chemical potentials at the same temperature and pressure. Different compositions at a common pressure can instead give different chemical potentials for one component through its thermodynamic activity.
Equality concerns the same exchangeable component in different places, not all different chemical species. For a permitted chemical reaction, the corresponding interior chemical equilibrium criterion is , with signed stoichiometric vector entries . The existing chemical equilibrium and constrained Gibbs minimization for chemical equilibrium sections develop those composition constraints.
Unconstrained thermal, mechanical and particle-exchange contacts require equality of temperature, pressure and the chemical potential of each exchangeable species. Walls, external fields, interfaces and chemical reactions change which variations are allowed.
Rigid compartments impose a different constraint: fixed volume rather than fixed pressure. With Helmholtz free energy ,This follows immediately by differentiating in the chemical potential differential above. It expresses the same chemical potential through the thermodynamic potential appropriate to fixed temperature and volume.
Consider a closed system consisting of two rigid compartments of equal volume , maintained at a common fixed temperature . A valve permits exchange of the same ideal gas; the partition stays in place. Initially and , with conserved total amount of substance . These are macroscopic moles, modeled continuously, not twelve individual particles. By the ideal gas law, , so initiallyAt with the stipulated , this is approximately , favoring transfer from A to B.
At fixed and equal , integrating the ideal-gas chemical potential gives the total Helmholtz free energy, up to a term independent of the allocation,Here is a positive reference amount of substance; any omitted common term linear in is constant along this exchange. Let be the dimensionless fraction in A. The relative Helmholtz free energy then takes the particularly simple formIts derivative along the allowed transfer and its second derivative areThus the Helmholtz free energy is a strictly convex function of the allocation for positive amounts. Its unique minimum has , or , and equal chemical potentials. Initially its excess above this minimum is at . Relaxation decreases the total Helmholtz free energy by this amount in the model; the final pressures are also equal because both compartments have the same volume.
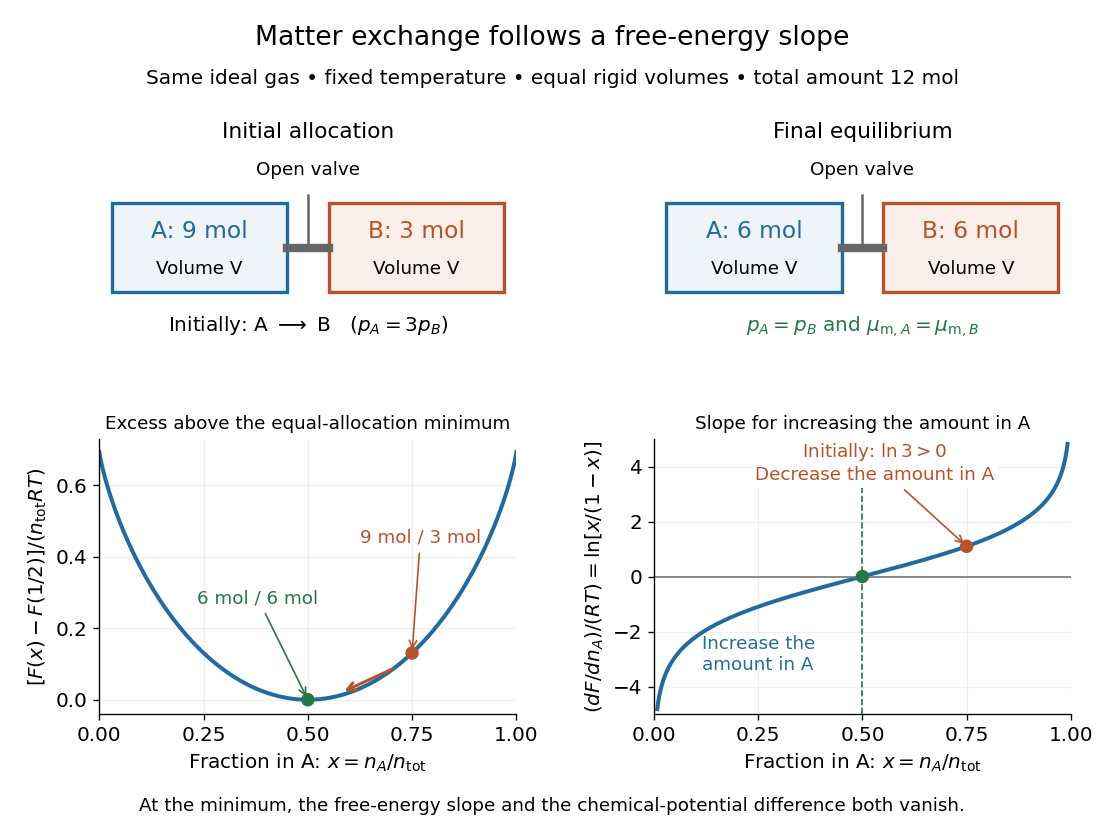
The beautiful connection is that the direction of matter exchange and the slope of a free energy curve are the same calculation. The entropy gained by redistributing the ideal gas makes equal allocation favorable here. As with chemical equilibrium, a free energy minimum identifies the favored state, while transport properties determine how quickly it is reached. The fixed-temperature, fixed-pressure Gibbs free energy criterion is summarized in 16.4 Free Energy; the rigid experiment instead uses Helmholtz free energy.
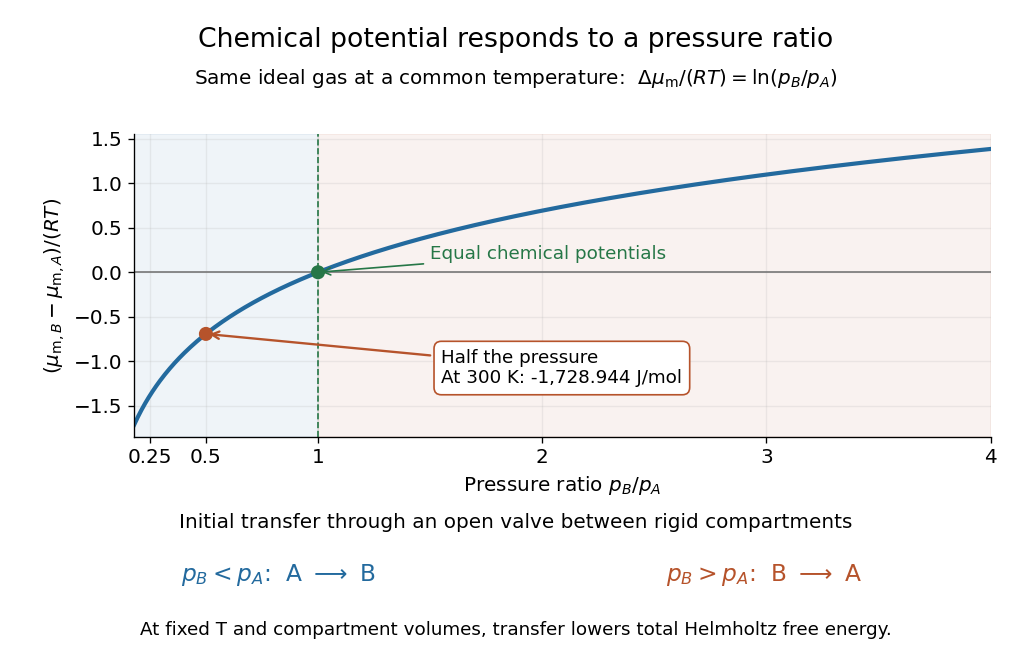
For a pure ideal gas at temperature and pressure ,where is the molar gas constant and specifies the reference pressure of the standard state. The ratio inside the natural logarithm is dimensionless. The chemical potential difference therefore depends on a ratio of pressures, not an absolute difference. See MIT 5.60, Lecture 14, page 4.
For a chemical species in an ideal gas mixture, replace by its partial pressure. More generally write using its thermodynamic activity and a consistent standard state. For an ideal dilute chemical solution, with reference concentration ; the thermodynamic activity section describes departures from this idealization.
As a computed comparison, take identical chemical species at , with and . Stipulate . ThenHalving the pressure gives the same decrement whether starting from or , provided the ideal gas approximation remains valid. This comparison uses different pressures; it is not the common-pressure reservoir setup of chemical potential equality for particle exchange. The fixed-volume example in chemical potential and Helmholtz free energy supplies an exchange experiment with unequal initial pressures.
The molar electrochemical potential combines the molar chemical potential with the electrical contribution , where is the signed charge number of an ion, is the Faraday constant and is electric potential. Both terms have units . In the particle convention the corresponding expression is , with elementary charge ; multiplying it by the Avogadro constant gives the molar form. Transfer between locations at the same temperature and pressure is driven by differences in electrochemical potential. The membrane transport examples use it to distinguish neutral solute transfer from ion transfer.
Define the displayed quantity as the free energy available per unit positive charge for sodium entry, with . Its negative is the signed inward electrochemical cost per charge. A reversal of the transport direction reverses the sign. At equilibrium it vanishes without requiring equal concentrations.
Start at sodium electrochemical equilibrium with internal ions. Exporting ions changes the membrane potential by and the internal number density to . Subtracting the initially balanced chemical and electrical contributions gives the displayed incremental cost. An initial zero potential further implies for an ideal fixed external reservoir. Nonzero initial equilibrium potential permits unequal concentrations, so that latter equality is an additional assumption.
An electrochemical gradient combines spatial changes in chemical potential and electric potential for a charged species. Passive transport proceeds downhill in electrochemical potential when an appropriate route is available; a concentration difference alone does not determine the direction for an ion.
A proton-motive force is a proton electrochemical potential difference per unit electric charge. It combines the electric potential difference and the chemical potential difference across a cell membrane; the direction and sign convention must be specified.
For a composition free energy , the local chemical potential is the functional derivativeAt phase coexistence, exchanges between the bulk phases require equal chemical potentials. Mechanical equilibrium additionally requires equal pressure for a flat interface, or a Young–Laplace equation pressure jump for a curved interface.
At chemical equilibrium, a reversible chemical reaction has equal forward and reverse reaction rates, so its macroscopic composition stays constant. Individual molecules can keep changing identity: the reaction has not stopped. This dynamic balance is the definition described by the [ref]. Equal reaction rates also need not mean equal concentrations. A small population converting rapidly can balance a large population converting slowly.
This distinction gives two useful questions. Which composition is favored at the specified temperature and other constraints? Thermodynamics answers through chemical potentials and an equilibrium constant. How quickly does a prepared mixture approach that composition? Chemical kinetics answers through reaction rates. Knowing the destination does not determine the journey's duration. A composition that barely changes during observation may reflect extremely slow chemical kinetics, so constancy alone does not establish chemical equilibrium.
Consider the idealized elementary chemical reactionAssume a well-mixed closed system at fixed temperature and volume, with first-order law of mass action rates in both directions and no competing chemical reactions. Write and for concentrations in . The total is conserved because each conversion replaces one molecule by one molecule. This conservation is special to the chosen stoichiometry.
The forward and reverse reaction rates, measured as conversions per unit volume per unit time, are and . HenceSetting this derivative to zero givesWith and , solving the ordinary differential equation yieldsAt , these are approximately and . Initially and . At chemical equilibrium, both are , while the net reaction rate is zero. The persistent exchange is visible in the lower panel even after the upper panel has nearly flattened.
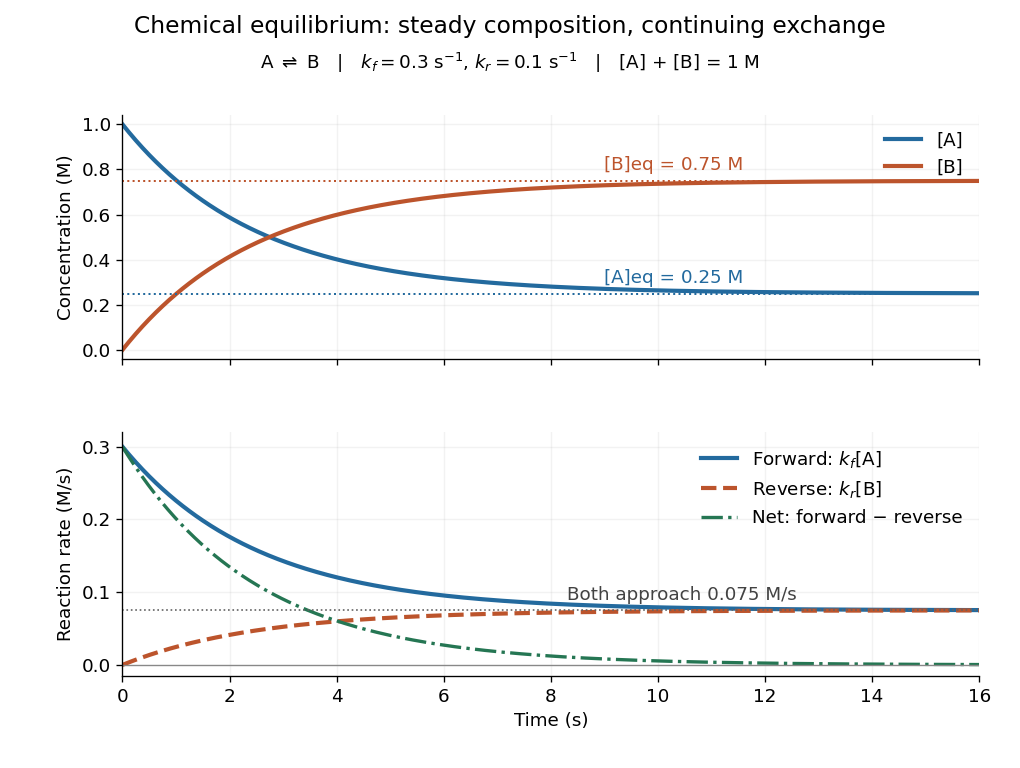
Constant equilibrium composition with continuing molecular conversion
. The idealized chemical reaction starts entirely in A. Upper panel: concentrations. Lower panel: forward, reverse and net reaction rates. The equilibrium rates coincide at a positive value. Details.Why does chemical equilibrium also involve a free energy minimum? For a closed system at fixed temperature and pressure, without imposed non-expansion work, stable equilibrium minimizes Gibbs free energy over compositions allowed by conserved elemental inventories. See W. C. Carter, MIT thermodynamics notes, sections 2-1.14 and 2-1.19. This is constrained Gibbs minimization for chemical equilibrium: freely minimizing over unrelated species amounts would change the inventory. At fixed temperature and volume, the appropriate potential is Helmholtz free energy. In either setting, varying an allowed extent of reaction gives the corresponding free energy slope , with molar chemical potentials and signed stoichiometric vector entries. An interior equilibrium has zero slope; stability requires a minimum. For A converting to B, this says .
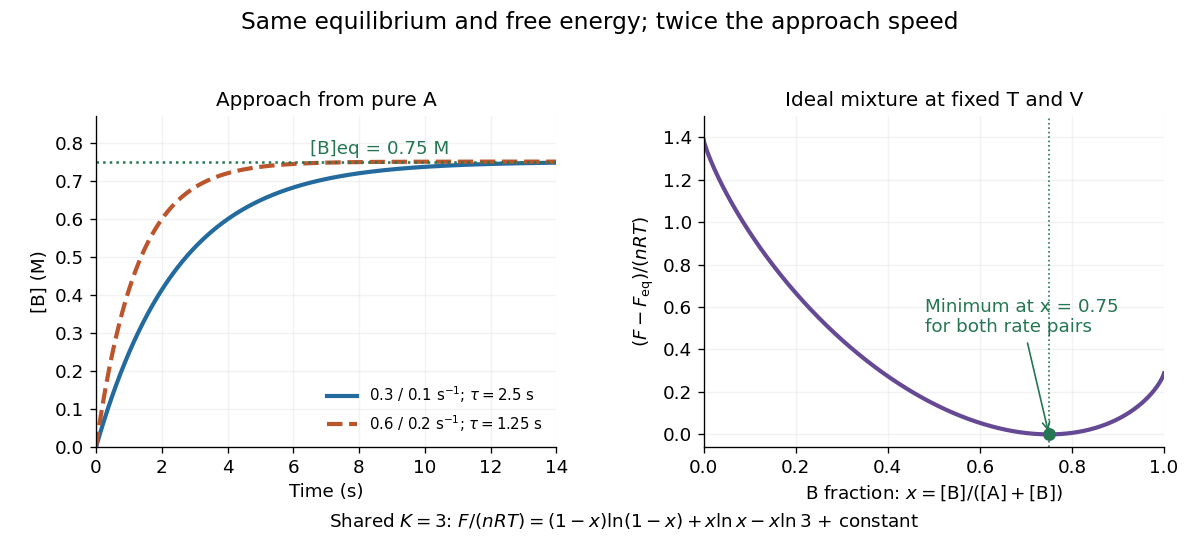
The beauty of this connection is that molecular exchange, a macroscopic composition, and a geometric minimum describe the same balance. In the ideal concentration model, let and choose the standard molar chemical potential difference consistent with . Up to a composition-independent constant, the fixed-volume Helmholtz free energy then obeyswhere is the conserved total amount of substance and the molar gas constant. Its derivative is , vanishing at . Its second derivative is inside , so this balance is the unique minimum. The mixing entropy terms explain why the favored composition still contains A.
For a mineral reaction producing one mole of water, let refer to the standard state and let the solids’ volume change be constant. If is the product/reactant solid-activity ratio, the Gibbs energy of reaction isFor temperature-independent specified activities and fugacity, equilibrium givesThe gas pressure work is already represented by the fugacity term; adding a second constant water-volume term double counts it. For an ideal gas, ; a given fugacity coefficient gives . If these quantities depend on temperature, the equilibrium equation must instead be solved implicitly. Pressure units in the solid volume correction must match those used for .
For ideal-gas single-particle functions and reaction , equilibrium gives . This particle-number ratio contains a volume factor; the concentration ratio does not.
For , the conserved amount is . The total particle count changes during reaction. Defining a dissociated fraction by gives .
For , . The left side increases monotonically on , so increasing the conserved formula-unit density at fixed temperature reduces dissociation in the ideal model.
The product of hydrogen-ion and hydroxide-ion activities at a specified temperature. In dilute aqueous solution at 25 degrees Celsius, its usual concentration approximation is ten to the minus fourteen.
The linked equilibria among dissolved carbon dioxide, bicarbonate, carbonate and hydrogen ions in seawater. At a fixed alkalinity, added carbon changes speciation and acidity, limiting how much extra carbon can be held at a given atmospheric partial pressure. Exchange and chemical partitioning therefore do not behave like irreversible destruction by a single first-order sink.
Adjustment of ocean and sediment carbonate chemistry through dissolution or burial of carbonate minerals. Following a carbon-dioxide perturbation, additional carbonate dissolution can increase alkalinity and permit further ocean uptake. The adjustment is much slower than surface gas exchange, operating over millennial timescales.
The temperature relation for a dimensionless thermodynamic equilibrium constant. With constant reaction enthalpy, integration gives the difference of reciprocal temperatures.
The derivative of Gibbs energy with respect to reaction extent at fixed temperature and pressure: . For a reversible electrochemical cell reaction, .
At fixed temperature, gives the pressure correction for the participating solids. Constant reaction volume reduces the integral to . A gas product treated by chemical fugacity instead contributes relative to its reference state. Adding a separate gas-volume correction to that term would double count the same pressure dependence.
The Gibbs energy of reaction for specified standard states. It determines the dimensionless equilibrium constant through .
For a nondegenerate spin-one-half ideal gas with energies , the two component chemical potentials are . Diffusive and spin-conversion equilibrium require one spatially constant common chemical potential, provided spin conversion is allowed. Hence the displayed densities give total number density and magnetization . The high-temperature fixed-density limit is , a Curie law. At fixed chemical potential the density itself varies with temperature, so the unnormalized magnetization need not have a pure dependence.
A chemical system is out of local chemical equilibrium when reaction adjustment cannot keep pace with transport, imposed changes or external driving. Comparing the chemical relaxation time with transport and irradiation times identifies plausible departures. Disequilibrium chemistry in an exoplanet atmosphere combines this principle with mixing, horizontal advection and atmospheric photochemistry.
For one particle species in chemical equilibrium under , chemical balance gives , hence . Elastic imposes no such constraint. In an expanding universe the number-changing reaction must remain fast compared with the Hubble parameter; after chemical freeze-out a nonzero chemical potential can develop even if elastic collisions maintain kinetic equilibrium.
The chemical relaxation time estimates how rapidly a perturbed abundance returns toward chemical equilibrium. Comparing it with the eddy mixing time identifies a chemical quench level.
For the A-to-B model in chemical equilibrium, deviations decay by exponential decay with . After one such interval, of the initial deviation remains; reaching one percent takes . Multiplying both rate constants by two preserves their ratio and the equilibrium composition, but halves to . The equilibrium exchange rates double to . Faster exchange therefore coexists with the same final composition and the same free energy minimum.
For a specified chemical reaction and standard state convention, the dimensionless equilibrium constant is the equilibrium value of the reaction quotient , where denotes thermodynamic activity and a signed stoichiometric vector entry. Thermodynamic activities incorporate the reference state and departures from ideal behavior. See IUPAC Gold Book, equilibrium constant.
In the ideal concentration model for A converting to B, use and with the same reference . Their ratio gives and . Equality applies to this elementary first-order pair; it is not a universal ratio of two measured rate constants for an arbitrary multistep chemical reaction.
At a specified constant temperature, with consistent standard states, the standard molar reaction Gibbs free energy isHere is the local slope per mole of extent of reaction at fixed temperature and pressure, rather than the finite total Gibbs free energy change between two mixtures. At equilibrium , while generally remains nonzero. For at , it is about . For positive activities, favors forward conversion and favors reverse conversion; neither sign supplies a reaction rate. See MIT 5.111, Lecture 18, reaction free energy and equilibrium.
For ideal concentration activities and signed stoichiometric coefficients , at equilibrium. Its numerical value depends on the stated standard concentration when . Number concentrations require a number-concentration reference; molar concentrations require the corresponding molar reference.
For , a concentration association constant is . Comparisons require matched solvent, temperature and standard-state conventions.
A standard state is a specified reference state used to define a species' standard chemical potential . The reference convention, including temperature, must be consistent when calculating thermodynamic activities, reaction quotients, and equilibrium constants.
The reference pressure used to define a standard gas activity; a common convention is one bar. Normalizing pressure by this reference keeps a thermodynamic equilibrium constant dimensionless.
The reference concentration used to make concentration activities dimensionless. Changing this reference changes the numerical dimensionless equilibrium constant when the reaction changes the total number of solute or gas molecules.
The thermodynamic activity is the dimensionless quantity defined by , where is the molar gas constant. For an ideal dilute solution with a specified concentration reference, ; departures from ideality are represented by an activity coefficient multiplying this ratio.
An effective gas pressure defined by in a specified standard-state convention. It approaches the gas pressure at low density and differs from the dimensionless fugacity.
The dimensionless ratio of a gas species' chemical fugacity to its pressure in the pure-fluid convention. Its thermodynamic activity relative to a gas reference pressure is . For mixtures the pressure denominator uses the species' partial pressure. The ideal-gas limit has .
For a chemical reaction with signed stoichiometric vector entries , positive for products and negative for reactants, the reaction quotient is the product of the species' thermodynamic activities raised to these powers. It equals the equilibrium constant at chemical equilibrium, with all quantities using the same standard state convention.
The chemical potential of photons in thermal equilibrium is zero because their number is not conserved.
Articles by others on the same topic



Chemical potential is a fundamental concept in thermodynamics and physical chemistry that describes the change in free energy of a system when an additional amount of substance is introduced, under constant temperature and pressure. It is a measure of the potential energy per particle in a system and reflects how the concentration of a species influences its behavior in a mixture.