Ionic bonding binds oppositely charged ions through attraction governed by electrostatics. In ordinary solid sodium chloride, each cation interacts with many anions, and also with other cations: attraction and repulsion together contribute to the binding of an extended ionic crystal. The chemical formula specifies equal numbers of the two kinds of ions; it does not divide the solid into isolated two-atom molecules.[ref]
This distinction explains a striking combination of properties: ordinary dry sodium chloride conducts electric current poorly, while its molten form or a sufficiently concentrated chemical solution in water conducts well. The ions already exist in the solid; their ability to move through the sample changes. A conductivity experiment on salt solutions tests this transport picture, rather than simply asserting that charged constituents must make every material conductive.
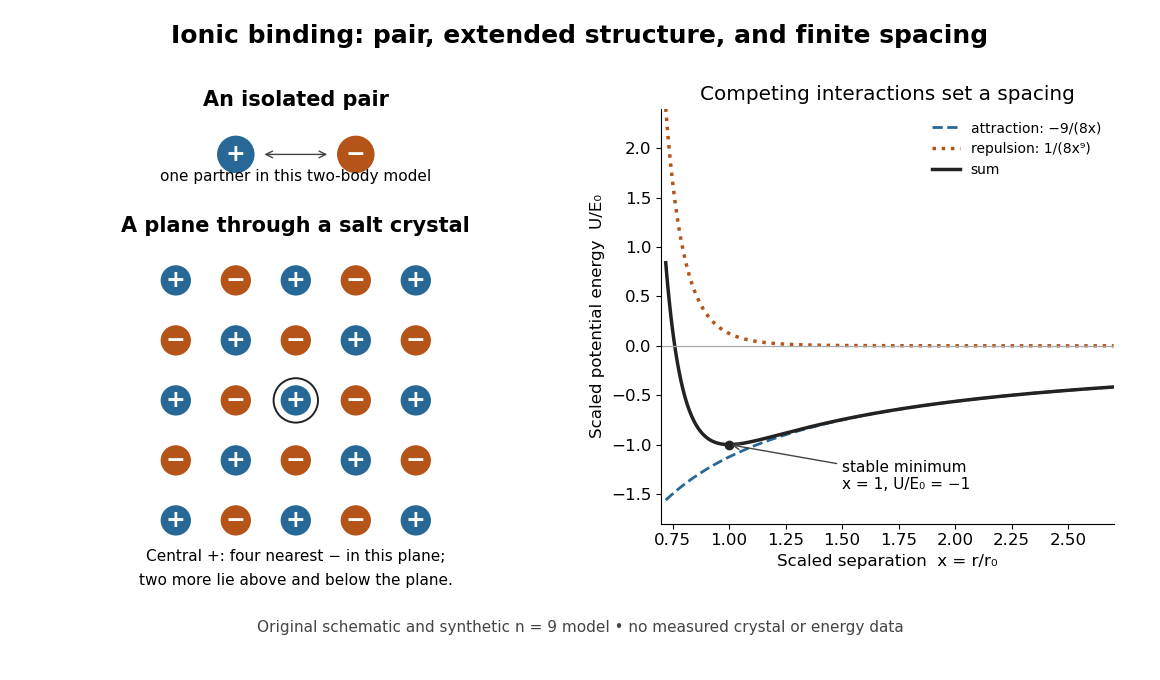
An isolated ion pair, a plane of an extended ionic crystal, and a synthetic potential energy model with a stable spacing
. Original schematic geometry and calculated model curves, not a measured electron density, diffraction pattern, or experimental lattice energy. Four nearest neighbours are visible in the plane of the rock salt structure; two more lie perpendicular to it. Disk sizes are arbitrary. The right-hand curves use the explicitly stipulated dimensionless model below. Details.For a large one-dimensional chain of alternating charges, is the Madelung constant. Summing the Coulomb interaction about one ion and dividing by two avoids double counting; per ion pair the result is . There are two nearest-neighbour bonds per pair, so a repulsion per bond gives the displayed second term. If , is a stable minimum with .
A hard-sphere model relating the radius ratio of opposite-charge ions to a possible coordination number. For a cube of eight mutually touching anions around a touching cation, ; real bonding and coordination-dependent radii limit its predictive power.
A geometric heuristic favouring the largest coordination number compatible with neighbouring opposite-charge ions touching the central ion without overlap. A limiting radius ratio rule comes from additionally making the surrounding ions just touch. It is a hard-sphere model, not an exact stability criterion.
A salt is an assembly of cations and anions with zero total electric charge. Its constituents can be single chemical atoms or groups joined internally by covalent bonds. Thus ionic bonding and covalent bonds can occur in the same chemical substance, at different structural levels.[ref]
Magnesium chloride is an ionic compound with two chloride anions per magnesium cation. Complete dissolution supplies one divalent magnesium ion and two monovalent chloride ions per formula unit. At analytical salt concentration , these species contribute molar ionic strength , provided additional dissolved ions are absent.
For a neutral formula unit, let count ions of type with electric charge , where is the elementary charge. The stoichiometry must satisfyFor and , the smallest positive counts are . For and , they are , giving calcium chloride : . The formula counts ions, not the number of nearest neighbours. A counterion is the oppositely charged partner needed to balance a charged species; its identity can change solubility and other properties without changing the identity of that species.[ref]
Historical formulation name; composition is identified by pharmacopoeial edition.
An ionic crystal repeats an arrangement of ions in three dimensions. A unit cell describes that repetition, whereas a formula unit describes composition. The two accounting devices answer different questions. Coulomb's law is not restricted to an arbitrarily drawn line connecting one cation to one anion: every ion contributes to the electric field experienced by the others.
The rock salt structure of ordinary sodium chloride has a coordination number of six for each kind of ion: six nearest neighbours of the opposite sign surround each ion along three perpendicular directions. Its conventional cubic unit cell contains four formula units; sharing corner, face, and edge sites between adjacent cells must be included in that count. Sixfold local coordination is compatible with overall stoichiometry.[ref]
The checkerboard plane in the diagram is a slice through this three-dimensional arrangement. It leaves out the two nearest neighbours above and below the plane. Repeating a complete even-by-even patch gives equal numbers of signs; the displayed odd-by-odd crop has a boundary imbalance and is not a complete neutral ionic crystal or a unit cell. Other salts need not have the rock salt structure or the same coordination number.
In this structural use, the coordination number counts nearest neighbouring constituents. State which neighbours are counted and which ionic crystal structure is being described; it is not an ion's electric charge number.
Eight nearest neighbours at cube corners around a central site; fluorite structure cations have this arrangement.
An ion pair is an associated cation and anion. An isolated ion pair lacks the extended neighbourhood of an ionic crystal. In a chemical solution, ion pairs can coexist with more widely separated ions; their abundance depends on conditions. A neutral gas-phase sodium chloride molecule can be treated as a discrete entity, with its own quantum mechanics and dissociation products. That use of molecule does not describe the constituents of ordinary solid sodium chloride.[ref]
The Feynman Lectures on Physics, Volume II by Richard Feynman and others. §8–3, distinction between vaporized NaCl pairs and separated gaseous ions
[ref]Oppositely charged ions remaining in direct contact without an intervening complete solvent shell. Their relative positions can bias substitution stereochemistry even when bond cleavage precedes nucleophile attack.
The bookkeeping steps and conserve electric charge and describe the electron inventories of separated species. They do not establish a chronological mechanism in which a labeled electron jumps between two pre-existing chemical atoms and becomes a classical object owned exclusively by one of them. In a bound material, quantum mechanics determines the electron density and the total energy.
Ionic bonding and a covalent bond are useful limiting descriptions with a continuum between them. Electronegativity helps anticipate unequal sharing of electron density, but a numerical cutoff does not measure a universal fraction of ionic bonding. A partial atomic charge depends on how the electron density is partitioned; integer formal charges are a bookkeeping convention. Using in a point-charge model therefore does not claim that every partition of the actual electron density assigns exactly those values.[ref]
The Feynman Lectures on Physics, Volume II by Richard Feynman and others. §30–2, “Chemical bonds in crystals”: gradations between covalent and ionic bonding
[ref]Chemistry 2e. §7.2, “Electronegativity and Bond Type”: general guide and exceptions
For two point electric charges in vacuum, Coulomb's law gives the interaction potential energywith zero at infinite separation. Here is the distance between them and is the vacuum permittivity. Opposite signs give negative potential energy and attraction; equal signs give positive potential energy and repulsion. For a finite collection of distinct point electric charges, sum over to count each pair once. A bulk ionic crystal requires the extended lattice sum, including both signs, rather than multiplying one attractive pair by the coordination number.[ref]
The Feynman Lectures on Physics, Volume II by Richard Feynman and others. §8–1, equations (8.1)–(8.3); §8–3, many-neighbour lattice sum
At short distances, overlapping electron density makes the simple point-charge picture inadequate. Quantum mechanics, including the Pauli exclusion principle, matters to the steep repulsive contribution. Pure attraction proportional to would collapse as ; it cannot set an equilibrium spacing on its own. A useful illustrative model isHere scales an assumed fixed arrangement, packages its attractive contribution, and represents short-range repulsion. The constants have units energy times length and energy times length to the power , respectively. This is a model, not a fitted description of a named salt.[ref]
The Feynman Lectures on Physics, Volume II by Richard Feynman and others. §8–3, close-distance repulsion and limits of a purely electrostatic lattice calculation
Differentiating gives and , so the stationary spacing is a stable minimum. For the diagram's stipulated , put and . ThenThe dimensionless curves show how competing interactions select a spacing. Their energy scale and exponent are synthetic; neither is a measured lattice energy or a melting prediction.
The Madelung constant, named for Erwin Madelung, is a dimensionless geometrical factor in the electrostatic binding of an ionic crystal. For a neutral lattice of electric charges with nearest unlike-ion distance , Coulomb's law gives the potential energy per ion pair as . At a positive reference site, , with neighboring electric charges . Both attraction and repulsion enter this lattice sum; summing with a neutral-bulk prescription is essential. Short-range overlap repulsion and other contributions must be added before inferring a stable spacing or a thermodynamic lattice energy.
A lattice energy needs a reaction, a sign convention, and a stated amount. Using the lattice-dissociation enthalpy convention for sodium chloride,The reverse lattice-formation enthalpy is negative with the same magnitude when initial and final states match. Chemistry 2e gives for this separation, per mole of sodium chloride formula units. This is about electronvolts per formula unit, not per individual ion. It concerns separated gaseous ions, rather than neutral chemical atoms, molten salt, or hydrated ions in water.[ref]
Chemistry 2e. §7.5, “Ionic Bond Strength and Lattice Energy”, positive convention, reverse convention and NaCl value
Also distinguish an internal energy change from an enthalpy change: . A static point-charge calculation omits motion and thermal contributions. Reversing a reaction reverses its energy change; replacing internal energy with enthalpy additionally changes the defined quantity. The Born-Haber cycle infers a lattice-formation enthalpy by combining other enthalpies, ionization energies, and electron affinities. These imagined stages are a thermodynamic accounting path, not a claim about the actual sequence of a chemical reaction.[ref]
A Born-Haber cycle uses Hess's law to infer a lattice energy from measurable enthalpy differences and isolated-species data. The motivating difficulty is that separating a mole of an ionic crystal into mutually separated gaseous ions is not a convenient direct calorimetry experiment. Instead, compare two accounting routes between the same elemental reference states and the same solid. Because enthalpy is a state function, the signed differences close exactly when all inputs describe compatible states. These routes do not assert a reaction mechanism or a temporal sequence of reaction intermediates.[ref]
For one mole of sodium chloride formula units, the overall formation reaction isThe stoichiometry requires half a mole of diatomic chlorine molecules, supplying one mole of chlorine chemical atoms. A half coefficient counts an amount of substance; it does not describe half of an individual molecule. Write each auxiliary chemical reaction with its own sign:Here is the sublimation enthalpy, the enthalpy of atomization of chlorine per mole of chemical atoms, and the molar ionization enthalpy represented by the ionization energy input in elementary treatments. If the bond-dissociation energy table gives the compatible molar enthalpy for , use half that number for . Both ions in the last step are gaseous, and no aqueous hydration of ions belongs in that step.
Adding these equations cancels the gaseous chemical atoms, the gaseous ions and the free electron, leaving precisely the overall formation reaction. ThusThe closed traversal returns from the solid to the elemental reference states, adding ; therefore . Notice that the signed attachment term is negative. Substituting a positive spectroscopic electron affinity for would reverse the attachment contribution.
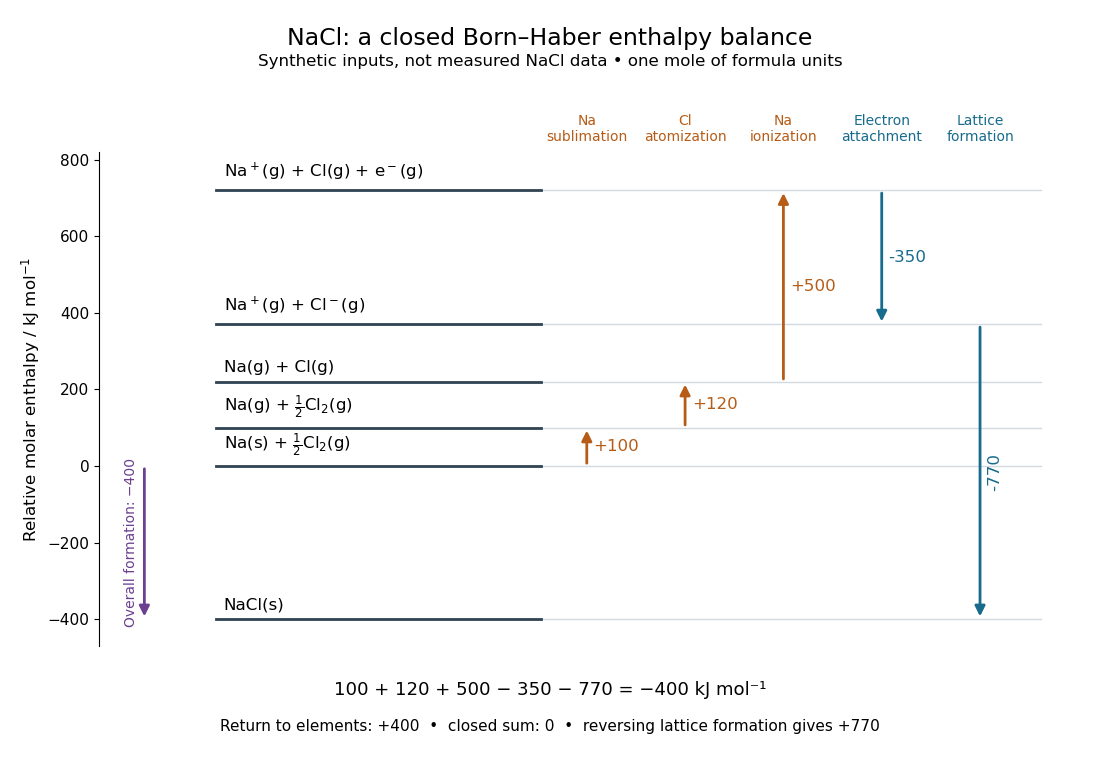
A closed Born-Haber cycle for sodium chloride using explicitly synthetic molar enthalpies: levels represent complete material inventories, and horizontal position is bookkeeping rather than a reaction coordinate
. Details. The diagram deliberately uses invented, internally consistent inputs in : , , , and . Preparation of separated gaseous ions raises the enthalpy by , so and . The closure check is . These numbers are not measured sodium chloride data and do not replace the source's example in lattice energy. Every level retains one sodium nucleus, one chlorine nucleus and zero total electric charge; the ionization level explicitly retains the free electron.
A second indirect route makes the experimental distinction especially clear: a calorimeter can measure dissolution of a weighed salt into a known amount of water, but its final ions are hydrated, not separated gaseous ions. At compatible standard states and the same temperature,Here the combined hydration of ions term takes one mole each of gaseous and to the specified aqueous reference state. Its sum suffices: dissolution calorimetry alone cannot independently assign single-ion hydration of ions enthalpies, or determine without another input. A finite-concentration measurement also requires dilution corrections before it can supply .
In a simple constant-pressure solution calorimeter, an insulated vessel, stirrer and thermometer follow the temperature change after mixing. A calibrated effective heat capacity converts the corrected temperature change to heat. With no applied heating, negligible external heat transfer, and only pressure-volume work, the approximate molar result isHere is the amount of substance dissolved and includes the chemical solution and the vessel. Cooling means positive dissolution enthalpy, because the dissolving sample takes heat from its surroundings.[ref]
Chemistry 2e. §5.2, Figures 5.11–5.13 and discussion of including solution and calorimeter heat capacities
An actual reference measurement used potassium chloride, not sodium chloride: a platinum-lined calorimeter with an adiabatic shield and a quartz-oscillator thermometer. Because dissolution absorbed heat, measured electrical heating prevented loss of shield control; the readouts were electrical input and the vessel/shield temperature record. Stirring was checked visually in a dummy vessel. The thermometer was compared with a calibrated resistance thermometer, and electrical calibration bracketed the reaction. The reported result was at , for one mole of potassium chloride in moles of water.[ref]
The Enthalpy of Solution of SRM 1655 (KCl) in H2O by Marthada V. Kilday. §3 and §4: apparatus, calibration, readouts and specified dilution; inspected through indexed article text
Checks on residual water, particle size and final concentration, and agreement with a second calorimeter, constrained systematic errors. Imperfect shield tracking required heat transfer corrections. This experiment measures an enthalpy of solution, not a direct lattice energy; the hydration and dilution bookkeeping still has to be supplied. The full article could not be opened here; the inspected indexed text supports the stated apparatus and checks, without reconstructing its complete uncertainty budget.[ref]
The Enthalpy of Solution of SRM 1655 (KCl) in H2O by Marthada V. Kilday. §§2–4 and conclusion: moisture, particle-size and dilution studies; confirmation with an isoperibol calorimeter; full-page access blocked
Many ionic crystals have high melting temperatures compared with small molecular solids, because reorganizing their strongly interacting ions costs energy. But melting leaves a dense liquid with interactions; it does not perform the complete separation defining a lattice energy. At fixed pressure, the transition occurs when the Gibbs free energies of the solid and liquid match. The relevant enthalpy and entropy differences both matter, so lattice energy alone is not a universal ordering of melting temperatures. Strong covalent bonds in extended networks can also give high melting temperatures.[ref]
Chemistry 2e. §10.5, “Ionic Solids” and “Covalent Network Solid”; §16.4, equations defining Gibbs free energy and its change
Solubility specifies the amount that can remain dissolved at chemical equilibrium under stated conditions; it is different from how quickly a sample dissolves. Separating ions from an ionic crystal costs energy, but solvation establishes new interactions with the solvent. In water, hydration of ions couples charged ions to polar water molecules. The net enthalpy of dissolution can have either sign; entropy and concentration also enter the Gibbs free energy balance. Consequently, being a salt does not guarantee high solubility in water or any other solvent.[ref]
Chemistry 2e. §11.1, “The Formation of Solutions”, Figure 11.4; §11.3, definition of solubility; §16.4, free energy
Low solubility must also be distinguished from weak electrolyte behavior. A small amount of an ionic compound can dissolve essentially completely as ions, giving few charge carriers because little material dissolved. Calling that a weak electrolyte solely because its chemical solution conducts poorly would conflate two different questions.[ref]
Solvation is stabilization through interactions between a solute and its solvent. It involves both the solute's surroundings and reorganization of the solvent: neighbors move, orientations change, and existing interactions compete with new ones. Electric forces, Van der Waals forces, and specific interactions such as hydrogen bonds can contribute. Solvation can also occur at groups on an insoluble material, so it does not require complete dissolution.[ref]
Solvation by water is called hydration. Hydration of ions is the charged special case, but neutral molecules such as sucrose can also interact with water without becoming ions. A hydrated cation retains its positive net electric charge; orienting neutral water molecules around it does not supply an extra electron. Dissolution, desolvation, and ionization describe different changes, as the examples below show.[ref][ref]
11.2 Electrolytes. Chemistry 2e by Paul Flowers and others; “Ionic Electrolytes” and “Covalent Electrolytes”
The useful question is therefore not just whether a solute attracts a solvent, but how much the whole system gains after paying for the rearrangements. Solvation free energy describes a specified transfer into a solvent; solvation thermodynamic balance explains why even cooling during dissolution can accompany favorable mixing. The existing solubility discussion concerns how much remains dissolved at chemical equilibrium.
A ratio of concentrations or activities between specified phases. State whether all protonation states are combined: a pH-dependent total distribution ratio differs from a partition coefficient for one molecular species.
Reorganization of a solvent to accommodate a solute. Its free-energy cost competes with favourable solute–solvent interactions and depends on solute size and shape.
A solvation shell is the region of solvent constituents surrounding a solute, with local arrangements influenced by their interactions. The first solvation shell contains the closest neighbors under a stated distance or interaction criterion. A coordination number counts those neighbors; it requires such a criterion and can fluctuate. Neither a drawn ring nor a few illustrated molecules determines a measured coordination number.
In a liquid, a solvation shell is an evolving neighborhood: solvent molecules reorient and exchange with the surroundings. Different ions can produce different local dynamics. Spectroscopic measurement of hydration dynamics provides an experimental way to distinguish some of those environments without pretending that a shell is a fixed cage.
A hydration shell is a solvation shell in water. The existing hydration of ions discussion describes preferential orientations around cations and anions.
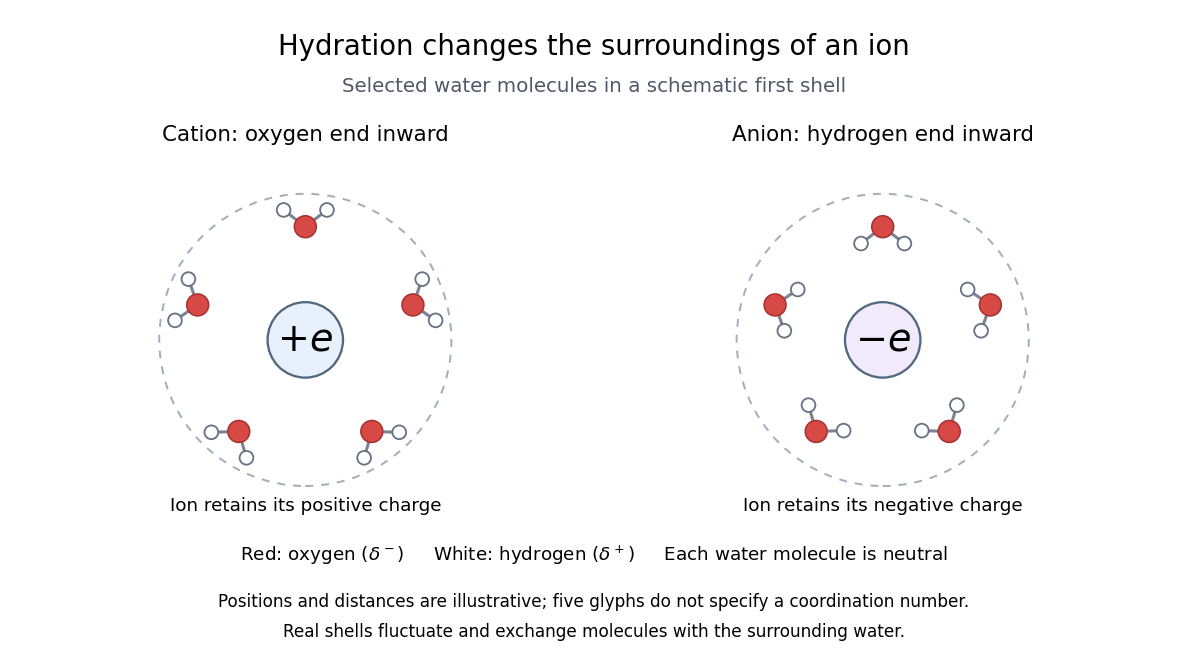
Qualitative hydration shells: opposite orientations of polar water molecules around a cation and an anion, with each ion's net electric charge retained
. Original schematic, not measured molecular positions. Atom spacings and the dashed shell boundary are illustrative. Five selected water glyphs in each panel do not assert a coordination number. Each water molecule is neutral despite its partial atomic charges. Details.An ion-dipole attraction couples an ion's electric field to a molecule's electric dipole moment. In the elementary point-electric dipole approximation, the interaction potential energy is . An orientation with the electric dipole moment along the electric field lowers this energy. For water, the oxygen end preferentially faces a cation, and the hydrogen end an anion. At nonzero temperature, fluctuations and competing hydrogen bonds prevent perfect alignment of every molecule.[ref]
11.2 Electrolytes. Chemistry 2e by Paul Flowers and others; “Ionic Electrolytes”, Figure 11.7 and its explanatory paragraph
What separates a hydration shell from the surrounding water experimentally? Electrical conductivity addresses a different question. The documented lamp demonstration immerses metal electrodes in a chemical solution, rinses them between samples, and records brightness; adding sodium chloride to distilled water makes it conductive. A water blank and dissolved sucrose comparison test whether dissolved material alone explains the signal. Fixed geometry, temperature, and stated concentrations are additional proposed controls. This reveals mobile electric charge, with limits from contamination, electrode interfaces and detection threshold; it supplies no shell positions. Conductivity experiment on salt solutions explains the apparatus and controls in detail.[ref]
Conductivity of Solutions by Harvard Natural Sciences Lecture Demonstrations. “Setting it Up” and “Comments”
In the authored study Influence of ions on the hydrogen-bond structure in liquid water by Anne Willem Omta and others, pump-probe spectroscopy uses tunable mid-infrared pulses traversing a sample. The probe's wave polarization is set at to the pump; after the sample, splitting and polarizers select parallel and perpendicular transmission. Measuring each transmission with and without the pump gives and , the respective logarithms of the transmission ratios. Their dimensionless anisotropy isIts decay probes reorientation rather than merely loss of vibrational excitation. Dilute semiheavy water in heavy water separates the O–H and O–D bands. Varying the semiheavy water concentration checked that transfer of excitation between nearby O–H groups did not determine the decay. These apparatus and control details are from the inspected author-uploaded article, not a new experiment.[ref]
The related thesis chapter documents two O–H absorption bands in a sodium perchlorate chemical solution: approximately for groups forming hydrogen bonds to water, and for groups forming hydrogen bonds to the anion. Selecting pump and probe bands, then a later delay window, separated their responses. The anion-associated anisotropy decayed more slowly; the water-bonded component showed no resolved change in decay time across the tested sodium perchlorate concentrations. The chapter also used O–D excitation to extend observation beyond the shorter O–H vibrational lifetime. These results support distinct local dynamics rather than a permanently fixed hydration shell. Spectral overlap, finite observation windows, and the selected salts limit the inference: this does not prove that every ion leaves all distant water properties unchanged.[ref]
Pump-probe spectroscopy measures a sample's response to a probe pulse at a controlled delay after an exciting pump pulse. Varying delay follows dynamics; selecting frequency and wave polarization can distinguish different contributions. Spectroscopic measurement of hydration dynamics is an application to water surrounding ions.
Semiheavy water contains one ordinary hydrogen nucleus and one deuterium nucleus. Dilute semiheavy water in heavy water provides relatively isolated O–H groups for infrared spectroscopy.
Dissolution transfers material into a chemical solution. It can include separation from an ionic crystal and solvation, or dispersal of neutral molecules. For sucrose and sodium chloride, respectively,Here specifies water as the solvent; it does not specify a fixed number of attached water molecules. The neutral sucrose remains molecular. Dissociation of an ionic compound separates pre-existing ions in the second example; solvation stabilizes their new surroundings.[ref]
11.1 The Dissolution Process. Chemistry 2e by Paul Flowers and others; Opening molecular and ionic dissolution examples
[ref]Ionization instead creates ions from neutral species. Hydrogen chloride reacting with water illustrates protonation of water rather than mere rearrangement of a hydration shell:The hydronium and negative ion together preserve total electric charge. Solvation can influence this chemical reaction, but an ion's solvation does not itself change its net electric charge.[ref]
11.2 Electrolytes. Chemistry 2e by Paul Flowers and others; “Covalent Electrolytes”, HCl proton-transfer example
Dissociation of an ionic compound separates its constituent ions. It differs from creating charged species by ionization of neutral molecules. Dissolved ions can also associate into ion pairs, so a dissociated description does not exclude all subsequent association.
Sodium perchlorate is the salt used in the spectrally selected example of spectroscopic measurement of hydration dynamics.
Desolvation removes some or all of the solvent environment of a species. Removing a hydration shell does not remove an ion's electric charge. Complete reversal of a specified solvation transfer reverses its Gibbs free energy and enthalpy changes when the same states and conventions are used. Partial desolvation, exchange of one solvent for another, and creation of a gaseous species have different endpoints; their costs need not match.
For transfer of the same species from a gaseous reference state into a solvent , define the molar solvation free energy byHere is a standard chemical potential. Negative solvation free energy means that transfer into the stated solvent lowers the Gibbs free energy relative to the stated gaseous reference. Include solute-solvent interactions, making room in the solvent, and its reorganization; counting only a favorable pair interaction omits part of the transfer.[ref]
State the temperature, gaseous reference pressure, dissolved reference concentration or other standard state convention, and species identity. Reference-state changes alter the quoted solvation free energy. For example, with an ideal dilute thermodynamic activity , replacing by while preserving the physical chemical potential requires to increase by . Thus the corresponding solvation free energy increases by if the gaseous reference stays fixed. This follows directly from , with the molar gas constant.
Solvent-specific stabilization means that the same species need not have the same solvation free energy in different solvents. Chemical polarity, hydrogen bonds, and the cost of reorganizing each solvent all matter. Water can stabilize ions by ion-dipole attractions; a favorable isolated attraction alone still cannot decide the total dissolution balance.[ref]
With a common gaseous reference and compatible dissolved standard states, subtraction removes that reference:A negative result favors solvent B under those reference conventions. This comparison assumes the same species : if protonation, ionization, or formation of ion pairs changes its identity, include a chemical equilibrium for each corresponding chemical reaction.
At fixed temperature and pressure, compare fully specified initial and final states usingThe entropy change is for the whole selected system, including both solute and solvent. Dispersing a solute can increase accessible arrangements while orienting nearby solvent molecules restricts others. One picture of an ordered hydration shell cannot determine the sign of the total entropy change. A negative Gibbs free energy change favors the specified process; it does not determine its reaction rate.[ref]
An illustrative enthalpy accounting path separates solute constituents, reorganizes the solvent, and establishes their interactions:This is a chosen decomposition between compatible states, not a measured sequence. If a gaseous-ion-to-chemical solution solvation term already includes solvent reorganization, do not add that reorganization a second time. Lattice enthalpy from solution calorimetry gives the alternative Hess's law route that combines lattice separation with the complete hydration of ions term.[ref]
11.1 The Dissolution Process. Chemistry 2e by Paul Flowers and others; “The Formation of Solutions”, Figure 11.4
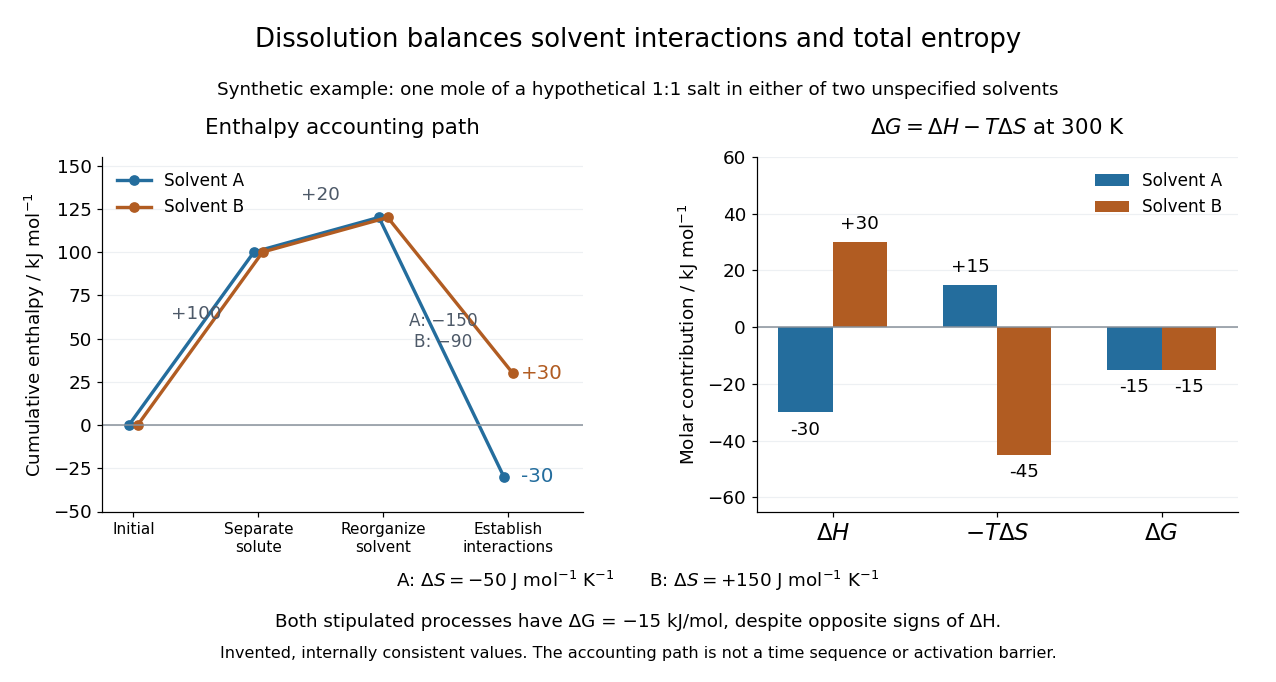
Synthetic dissolution bookkeeping: solvent interactions change the enthalpy balance, while total entropy also enters the Gibbs free energy
. Original stipulated model for one mole of a hypothetical 1:1 salt at , with two unspecified solvents. No plotted value is a measurement or a claim about a named chemical substance. The accounting path is not a reaction coordinate or an activation energy. Details.The invented inputs are for separation and for solvent reorganization, in . Interaction terms of in solvent A and in solvent B give and . Stipulated total entropy changes of and give and at . Both selected processes therefore have . This deliberately constructed comparison shows why the signs of the enthalpy and Gibbs free energy changes need not agree; it does not predict either solvent's solubility.
For incremental dissolution at a particular composition, the relevant chemical potential difference also depends on thermodynamic activities. At chemical equilibrium that driving difference vanishes. A quoted standard Gibbs free energy change and the finite change between arbitrary mixtures answer different questions; use compatible standard states and the reaction quotient when discussing solubility.
The motivating calorimetry experiment is especially instructive. A weighed potassium chloride sample dissolves in a known amount of water in a platinum-lined, shielded calorimeter; a quartz-oscillator thermometer and measured electrical heating provide the readouts. Electrical calibration before and after the reaction, comparison with a resistance thermometer, and checks on residual water and particle size constrain errors. The inspected authored article reports positive enthalpy of solution at for the specified dilution, despite complete dissolution. This is a combined enthalpy balance, not an isolated solvation free energy or direct single-ion hydration of ions measurement. Shield tracking and final concentration matter; the existing lattice enthalpy from solution calorimetry discussion gives the numerical result and the conversion limits.[ref]
The Enthalpy of Solution of SRM 1655 (KCl) in H2O by Marthada V. Kilday. Abstract, §§3–4 and conclusion; inspected indexed authored text, direct PMC access challenged
Hydration of ions is solvation by water. The polar water molecules orient preferentially around cations and anions, though the surrounding arrangement fluctuates rather than forming a permanently fixed cage.[ref]
The enthalpy change on transferring a specified gas-phase ion to the specified aqueous standard state. Charge density strongly affects the stabilization; single-ion values require a consistent convention.
How can a clear chemical solution reveal otherwise invisible mobile ions? The documented Conductivity of Solutions by Harvard Natural Sciences Lecture Demonstrations demonstration uses metal electrodes enclosed in tubing, an electrical supply, and a lamp in the circuit. The protocol specifies about samples in labeled beakers and a separate deionized-water rinse station. The recorded readout is lamp brightness after the electrodes enter each sample; the simplified variant tests water before and after adding sodium chloride. These are apparatus and procedure specifications, not new measurements reported here.[ref]
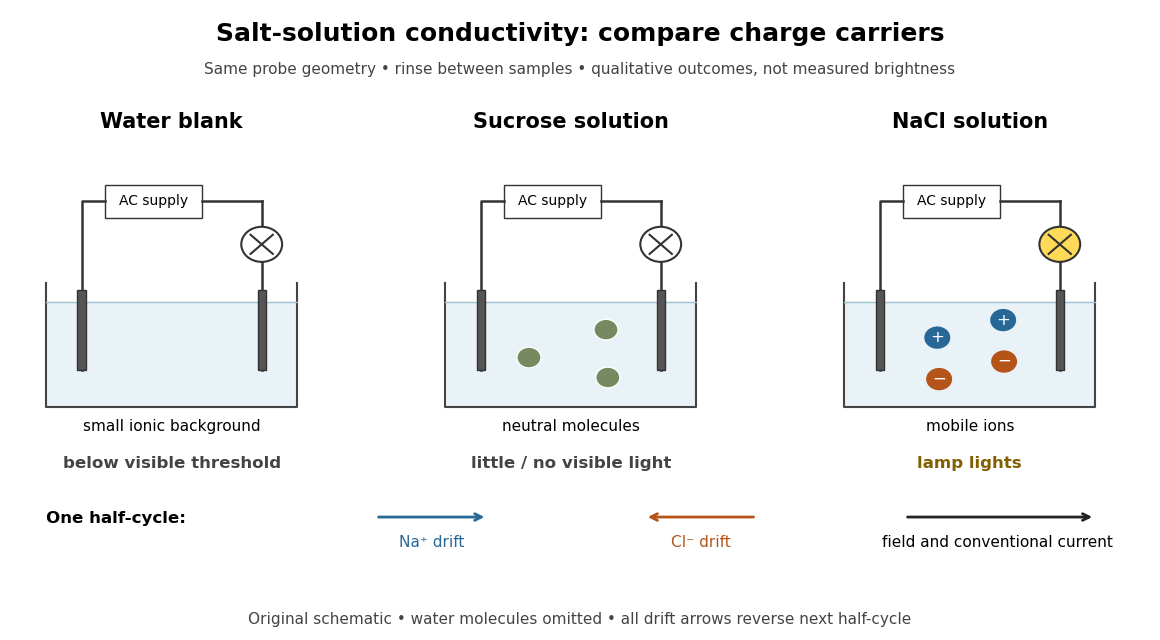
Electrical conductivity comparisons with a lamp tester, and the opposite motions of cations and anions in an electric field
. Original circuit schematic and qualitative model outcomes, not photographs or recorded brightness data. A dark lamp means a signal below this tester's visible threshold, not identically zero electrical conductivity. Arrows show drift velocities during one half-cycle of an alternating supply; both signs contribute to the same conventional current. Water molecules and hydration of ions are omitted from the microscopic panel. Details.The useful comparison is water, a sucrose chemical solution, and a sodium chloride chemical solution with the same probe geometry. A documented university account reports no visible light with water, a glowing lamp with dissolved sodium chloride, and a weak filament glow with its sucrose sample. The diagram summarizes the latter as little or no visible light; the precise detection threshold depends on the tester. Sucrose is a control for the idea that any dissolved material must conduct: its dispersed neutral molecules do not provide the abundant mobile ions supplied by the salt. This contrast is a documented teaching observation, not a numerical calibration of lamp brightness.[ref]
Chemistry 2e. §11.1, sucrose dissolves as molecules; §11.2, Figure 11.6 and ionic versus nonionic charge carriers
[ref]Rinsing the electrodes between samples controls carryover of dissolved ions, and the water blank checks the solvent background. Repeating the blank after the salt sample would test whether that rinse was effective. Additional methodological controls are fixed immersed area, electrode separation, supply setting, temperature, and comparison at stated concentrations. These additional controls are proposed checks, not undocumented claims about the source demonstration.
The inference is that mobile electric charge carriers complete the circuit. In the chemical solution, an electric field drives positive ions along the field and negative ions against it; multiplying each drift velocity by its electric charge makes both contributions to conventional current point the same way. With an alternating supply the direction reverses each half-cycle. The external wires instead carry electric current through electrons. The lamp alone identifies neither the species nor an exact electrical conductivity.
Important limits remain. Water has a small intrinsic population of hydronium and hydroxide, so a dark lamp does not prove absence of ions. Changes in probe geometry affect measured electrical conductance; electrolysis, electrode interfaces, contamination, and heating can also affect a real circuit. The comparison supports a transport model but does not independently measure a coordination number, a partitioned partial atomic charge, or a lattice energy.
Nor does poor conduction by ordinary dry sodium chloride establish that all ionic crystals are electrical insulators. Crystallographic defects can allow transport, and solid electrolytes can have substantial ionic electrical conductivity without melting. The authored abstract of A lithium superionic conductor by Noriaki Kamaya and others reports room-temperature ionic conduction in a solid electrolyte; the full experimental article was not accessible here.[ref]
A solid electrolyte carries electric current through mobile ions while remaining a solid. The distinction between ionic and electronic electrical conductivity matters: different charge carriers serve different roles in electrochemistry.
Zirconia stabilized by addition of yttria. At suitable elevated temperatures, oxygen vacancies allow oxide-ion transport, making the material useful in an oxygen concentration cell. Its oxide-ion transference number must remain sufficiently close to one for a simple ionic Nernst equation interpretation.
Articles by others on the same topic



Ionic bonding is a type of chemical bond that occurs when atoms transfer electrons from one to another, resulting in the formation of charged particles known as ions. This transfer typically occurs between atoms of significantly different electronegativities, such as metals and non-metals.