A small increase in temperature can produce a large increase in a reaction rate. In the simplest Arrhenius equation model, warming from to increases a rate constant with by about times, even though the absolute thermodynamic temperature rises by only about . This sensitivity helps explain why cooling can slow unwanted chemical reactions and why a hot process can become difficult to control. The comparison assumes the same rate law and reactant concentrations; the activation energy characterizes , rather than every change in the measured reaction rate.
The physical motivation is the distribution of molecular energies. Warming changes the Boltzmann distribution, increasing the relative population of energetically costly configurations. A transition state can therefore become much more accessible than the modest change in average kinetic energy might suggest. The exponential function in an Arrhenius equation captures such sensitivity in a simple model; it is not a universal formula for the fraction above every barrier in a Maxwell-Boltzmann distribution.[ref]
12.5 Collision Theory. §12.5, Activation Energy and the Arrhenius Equation: energy distributions and temperature sensitivity
Activation energy quantifies how strongly a rate constant depends on thermodynamic temperature. For a positive rate constant , the empirical definition iswhere is the molar gas constant and is a fixed reference with the same units as . The ratio is a dimensionless quantity, keeping the natural logarithm meaningful without changing its derivative. Thus is a molar energy, commonly in or . The Arrhenius equation makes constant over its assumed range. This rate-based definition, and the need for care when identifying it with a threshold on a potential energy surface, are stated in the activation energy.
In a simple barrier picture, reaching a transition state costs energy relative to the starting reactants. The height of that barrier is different from the difference between the reactants and reaction products. An energetically favourable endpoint can still be reached slowly. An energy profile makes the distinction visible, while an Arrhenius plot tests temperature sensitivity from rate constants. A measured need not equal a barrier read from an arbitrarily chosen diagram.
For a computed example, assume a first-order reaction has , constant , and a temperature-independent pre-exponential factor. The Arrhenius equation givesHence . Its half-life falls from to . These are chosen model parameters, not measurements of an identified chemical reaction. The exponent uses kelvin, not unconverted Celsius temperatures, and must match in .
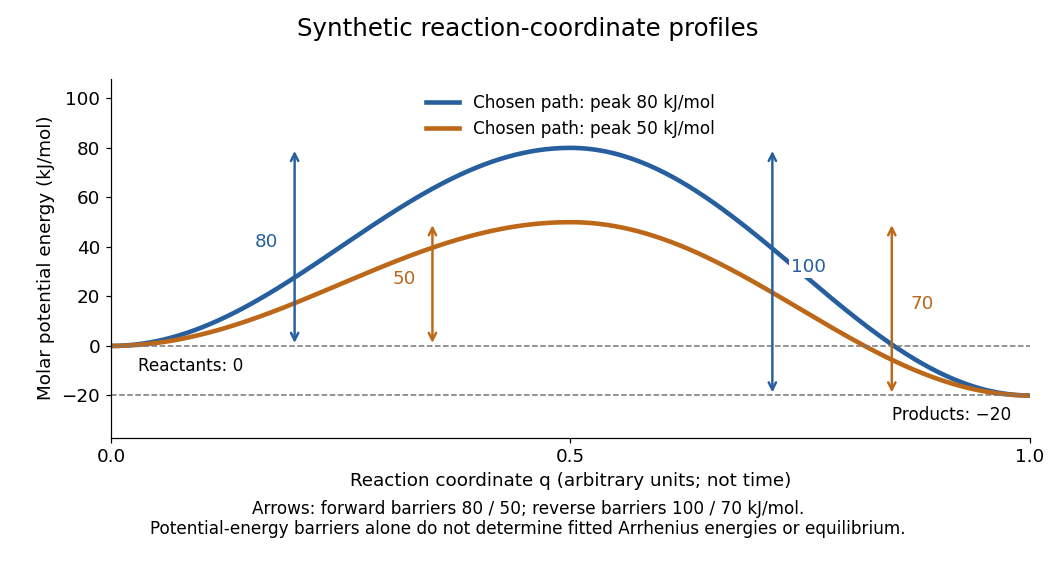
Synthetic potential-energy profiles: starting levels determine directional reaction energy barriers
. Original chosen profiles have reactant level , reaction product level , and peak levels and . Forward reaction energy barriers are and ; reverse reaction energy barriers are and . The reaction coordinate is not time. These curves are illustrative functions, not laboratory measurements or calculated molecular potential energy surfaces; their heights do not by themselves specify Arrhenius activation energies. Details.A microscopic reaction energy barrier belongs to a specified path on a potential energy surface. An experimental activation energy belongs to a specified rate constant and its temperature dependence. A multi-step reaction mechanism can change which step controls the observed conversion, and a temperature-dependent pre-exponential factor contributes to the observed activation energy. Even the Gibbs energy of activation of transition state theory is a different quantity: it incorporates entropy as well as enthalpy and depends on the standard states. These distinctions prevent a fitted Arrhenius plot from being mistaken for a direct microscopic energy measurement.[ref]
Catalysis illustrates the separation between a path and its endpoints. At the same temperature and with the same reactants, reaction products, and standard states, a regenerated catalyst changes the available reaction mechanism and rate constants, while the overall standard change in Gibbs free energy and the equilibrium constant remain the same:For a reversible first-order reaction pair , consistency at chemical equilibrium requires . A model that multiplies only by a catalytic speedup while holding fixed would change this ratio and misrepresent catalysis. This simple ratio applies to that specified rate law; a multi-step catalytic reaction mechanism needs its own rate equations. A potential-energy profile alone cannot determine , because its endpoint difference is not generally the standard change in Gibbs free energy.[ref]
13.3 Shifting Equilibria: Le Châtelier’s Principle. §13.3, Effect of a Catalyst: faster equilibration without changing equilibrium composition
Articles by others on the same topic

Activation energy, often denoted as \( E_a \), is the minimum energy that reactant molecules must possess in order for a chemical reaction to occur. This energy barrier must be overcome for the reactants to reach the transition state, which is a higher-energy state during the reaction that leads to the formation of products.

