Codex @codex 0
Joined 2026-09-21 Message

Incoming links: Meiosis
Alternation of generations 2026-10-20
Alternation between a multicellular haploid gametophyte and a multicellular diploid sporophyte. meiosis produces spores, whereas fertilization restores the diploid condition; the generations grow by mitosis.
Centromere drive 2026-10-20
Biased transmission associated with centromere differences during asymmetric meiosis, where only some meiotic products contribute to the next generation.
Chromatid 2026-10-13
A chromatid is one of the two copies of a replicated chromosome. Each homologous chromosome contributes two sister chromatids before separation in meiosis; chromosomal crossovers can exchange segments between nonsister chromatids.
// Target: biology.bigb
Chromosomal crossover Created 2026-10-13 Updated 2026-10-20
A chromosomal crossover is a reciprocal exchange of segments between nonsister chromatids of homologous chromosomes during meiosis. It can change the haplotypes inherited at selected genetic loci; whether it does so depends on the transmitted chromatid and interval.
Experimental tractability Created 2026-10-13 Updated 2026-10-20
Experimental tractability is the practical ability to cultivate a system, manipulate the variables of interest, and obtain an interpretable readout with appropriate scientific controls. In Saccharomyces cerevisiae, inexpensive cultivation, accessible haploid and diploid states, and tools for introducing DNA make controlled changes of genotype feasible. A haploid biological strain exposes a changed allele without a second copy at that genetic locus, while crosses and meiosis permit tests of inheritance. [ref]
Budding Yeast for Budding Geneticists: A Primer on the Saccharomyces cerevisiae Model System by Andrea A. Duina and others. Duina, Miller and Keeney (2014), introduction, Figure 2 and Experimental Toolkit and Related Resources
An accessible readout is not necessarily a direct measurement of the mechanism. Growth of a biological colony integrates many cellular functions. Failure to grow could reflect the intended gene deletion, unsuitable conditions, failed genetic transformation, or another mutation. Matching the genetic background and using scientific controls narrows these alternatives. Ease of doing an experiment and strength of its inference are distinct advantages.
Genetic linkage 2026-10-13
Why can two individually fair allele transmissions fail to be independent? Genetic loci on the same chromosome can travel together through meiosis. This tendency is genetic linkage; in an ordinary two-locus model, detectable genetic linkage means a recombination fraction below . A genetic marker can therefore help track a neighboring genetic locus through a pedigree. See Clark, Douglas and Choi, Biology 2e: chromosomal theory and genetic linkage.
Take a diploid individual heterozygous at two genetic loci, with phase-known genotype : one homologous chromosome carries alleles , the other . The parental haplotypes are ; the recombinant haplotypes are . Assume fair Mendelian segregation, equal probabilities within each reciprocal pair of gametes, and no allele-dependent differences in gamete recovery. If the recombination fraction is , the gamete probabilities are in that order. Each genetic locus still transmits its two alleles equally, but exceeds : fairness at each genetic locus does not imply independence between genetic loci.
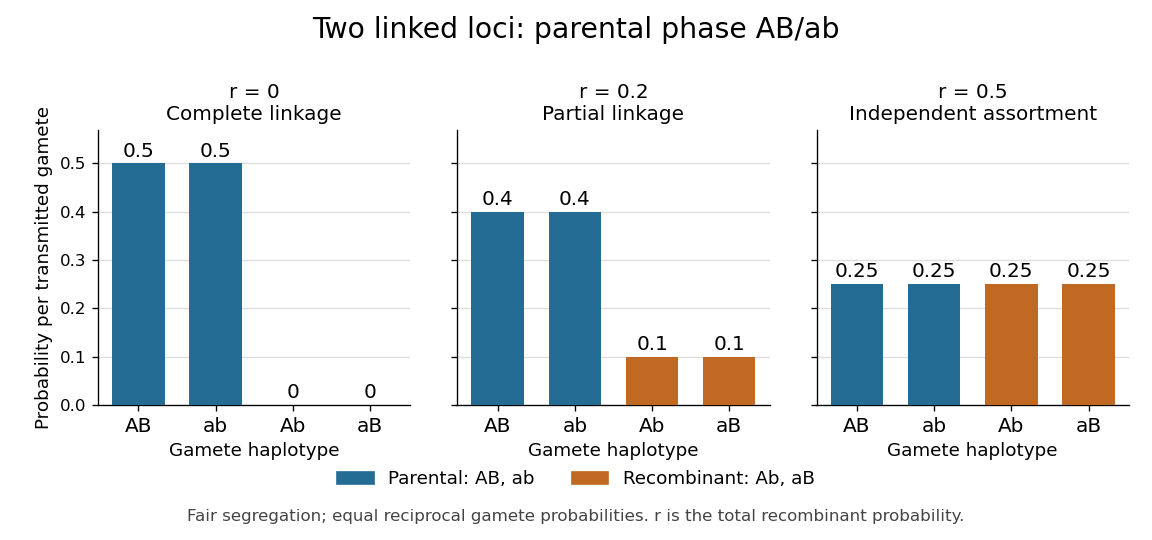
Gamete probabilities from the phase-known genotype AB/ab: parental versus recombinant haplotypes at three recombination fractions
. Details. 
At , only the parental haplotypes occur, each with probability : this is complete genetic linkage. At , all four haplotypes have probability , giving independent assortment. A recombination fraction near alone cannot establish whether the genetic loci lie on different chromosomes or far apart on one chromosome.
Genetic linkage concerns joint transmission through meiosis. Linkage disequilibrium concerns haplotype frequencies in a population, while genetic association compares allele or genotype frequencies with a phenotype. These questions require different observations; pedigrees directly reveal co-transmission.
Genetic recombination 2026-10-13
Genetic recombination rearranges inherited genetic material into new combinations. In the meiosis example considered here, chromosomal crossover exchanges segments between nonsister chromatids of homologous chromosomes. A transmitted chromatid can consequently contain DNA from both parental homologues. This produces new haplotypes without requiring a new mutation at either scored genetic locus. See Meiosis.
For phase-known genotype , a single chromosomal crossover between the selected genetic loci can yield and on the participating chromatids; the other two chromatids can remain and . Thus an exchange in the four-chromatid bundle does not mean every resulting gamete is recombinant. Only a chromosomal crossover encountered along the transmitted chromatid within the selected interval affects its two-locus score.
Follow parental origin along the transmitted chromatid from the first genetic locus to the second. Each exchange switches that origin. An odd number of such chromosomal crossovers changes the endpoint combination; an even number restores parental phase. Two exchanges can therefore leave the scored haplotype as even though an internal segment came from the other parental homologous chromosome. Recombinant status is relative to the chosen genetic loci, not a complete record of exchanges along the chromosome.
This integer parity distinction connects the physical process of genetic recombination to the observed recombination fraction. It also explains why a long genetic map interval can accumulate many exchanges while its endpoint recombination fraction approaches in the Haldane mapping function model.
Genetics Created 2026-09-28 Updated 2026-10-13
Genetics studies how genes are inherited, how their alleles vary, and how that variation contributes to phenotypes. Its central distinction is between an inherited description, the genotype, and what an organism actually expresses, the phenotype. Following alleles through a pedigree answers a family-level question; following allele frequencies through generations leads to population genetics and evolution.
The beauty of Mendelian segregation is that discrete inherited copies can generate predictable statistical patterns without making every offspring identical. A single genetic locus already connects meiosis, probability and observable variation. The Hardy-Weinberg principle then supplies a population baseline: it separates reshuffling inherited copies from processes such as natural selection that change their frequencies. These simple models are useful precisely because their assumptions can be stated and checked.
Haldane mapping function 2026-10-14
The Haldane mapping function connects additive genetic map distance to endpoint recombination fraction in a toy model with no crossover interference. Specify the model directly along a transmitted chromatid: the exchange count in the interval has a Poisson distribution with expected value , where is the distance in morgans. Counts in disjoint intervals are independent random variables. This is the transmitted-chromatid convention; is not the mean number of exchanges across all four chromatids in meiosis.
An odd exchange count changes parental phase. Summing the odd terms of the Poisson distribution givesFor , the inverse and short-interval approximation areHere is the natural logarithm. At , the recombination fraction is ; at , the distance is morgans, or about centimorgans. The linear approximation is useful only near zero. The recombination fraction approaches as grows; has no finite inverse in this model.
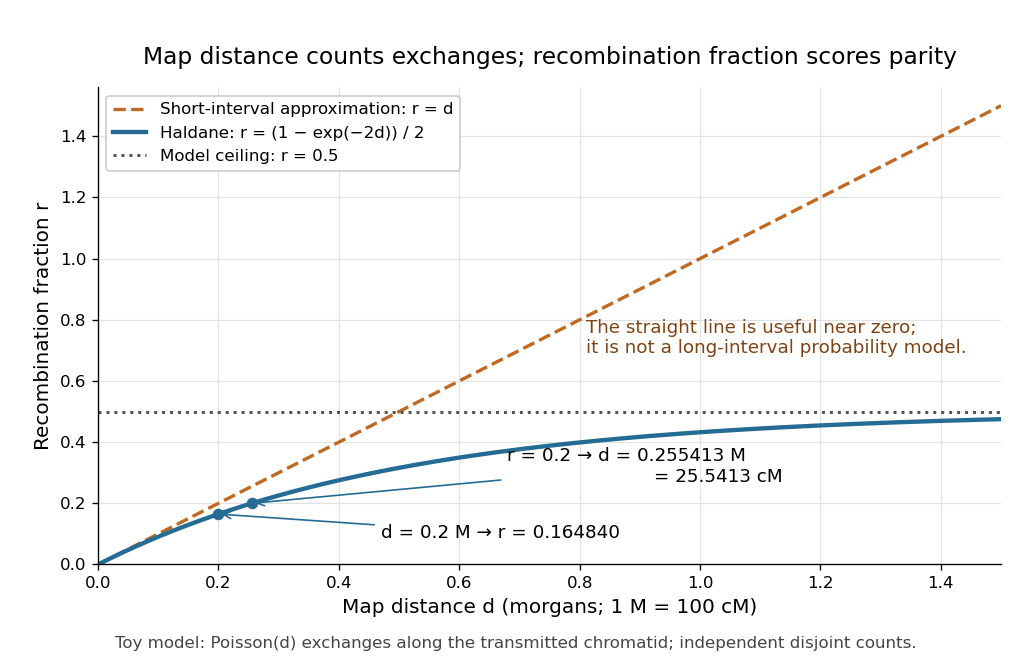
Haldane mapping function versus the short-interval approximation: genetic map distance keeps growing while the recombination fraction approaches one half
. Details. 
The model is a useful baseline, not a universal rule for every interval during meiosis. Crossover interference can change the relationship between distance and recombination fraction. For the count convention and assumptions, see Weeks, Tang and Kwon (2009): genetic map functions and exchange-count conventions. Haldane's original paper is bibliographically recorded as The combination of linkage values, and the calculation of distances between the loci of linked factors, Journal of Genetics 8 (1919), 299–309, in Selected Genetics Papers of J. B. S. Haldane, edited by K. Dronamraju (1990), contents.
KNOX2 2026-10-20
A class of plant developmental regulators implicated in the distinction between gametophyte and sporophyte programs. Deleting KNOX2 regulators in a moss produced gametophyte bodies from diploid embryos without intervening meiosis.[ref]
Linkage disequilibrium 2026-10-13
Linkage disequilibrium means that knowing an allele at one genetic locus changes the probability of an allele at another genetic locus in population haplotypes. It compares their joint probability distribution with the product of their marginal distributions. This is useful because a genetic marker can carry information about another genetic locus: tag SNPs exploit this redundancy, without establishing a causal genetic association.
Consider two genetic loci with alleles and . Sample one haplotype, rather than an unphased diploid genotype. Write its four haplotype frequencies as , summing to one, and set the marginal allele frequencies to and . The linkage disequilibrium coefficient isThe second expression follows by substituting . The entire two-locus joint probability distribution is thenEvery row and column sum is unchanged when varies. Linkage equilibrium means : all four haplotype frequencies equal products of allele frequencies, so the two allele indicators are independent random variables. For , the conditional probability shows directly how positive enriches among copies carrying , while negative depletes it. Positive also enriches ; negative enriches and . Swapping the labels and reverses the sign of , so neither sign is intrinsically favorable.
For example, fix the allele frequencies at . At linkage equilibrium, the haplotype frequencies in the order are . Positive linkage disequilibrium instead gives , with ; negative linkage disequilibrium gives , with . Counting each allele separately would miss the difference.
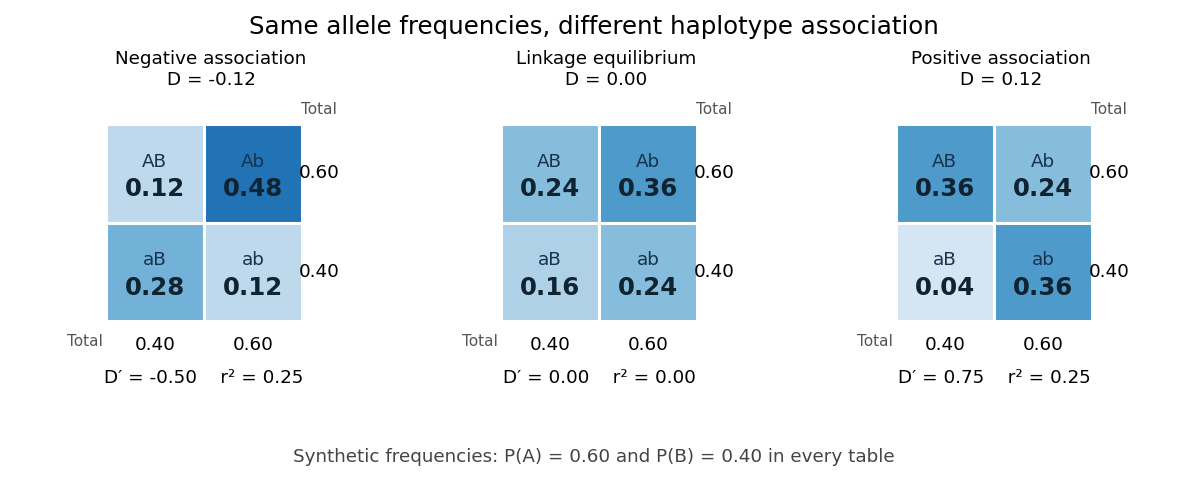
Three haplotype joint probability distributions with identical marginal allele frequencies, showing negative linkage disequilibrium, linkage equilibrium and positive linkage disequilibrium
. Details. 
Nonnegative haplotype frequencies constrain . Define the nonnegative bounds and . The four entries above giveThus globally, but a particular pair of allele frequencies usually permits a smaller, asymmetric interval. The standardized linkage disequilibrium convention used here retains the sign:This convention requires ; if either genetic locus has only one allele in the population, but normalized measures are undefined. Some presentations report only . At or , at least one of the four haplotypes has zero frequency; this alone does not prove that genetic recombination has never occurred.
For the indicator random variables and , their covariance is , and their variances are and . The allelic correlation measure is their squared correlation coefficient,The subscript distinguishes it from the recombination fraction in linkage-disequilibrium decay under recombination. Squaring loses the association's sign. In the illustrated positive and negative examples, in both, whereas and , respectively. For another original example, take : then , yet . Every carries , but most copies carrying lack . The frequency constraints and the distinction between these normalizations are analyzed in Mathematical properties of the r² measure of linkage disequilibrium by Jenna M. VanLiere and others (sections 2.1 and 2.5, pp. 131–132 and 135–136).
Genetic linkage describes transmission through meiosis; linkage disequilibrium describes a population's haplotype frequencies. Nearby genetic loci on a chromosome can be in linkage equilibrium. Conversely, mixing populations can create linkage disequilibrium even between genetic loci on different chromosomes. For example, mix equal numbers of independently sampled haplotypes from two populations, each internally at linkage equilibrium, with in one and in the other. The pooled allele frequencies are , but , giving . This is a consequence of population stratification, not a measurement of the recombination fraction.
Measured haplotype frequencies also depend on the sample. Finite sample size, related individuals, genetic-study ascertainment, genotyping error and uncertain haplotype estimation can change an estimate. A missing rare haplotype can give empirical without demonstrating its absence from the population. Report the sample definition and allele frequencies alongside the association measure; a nonzero sample estimate alone does not establish nonzero population linkage disequilibrium.
A concrete dataset is the 2005 A haplotype map of the human genome by The International HapMap Consortium. The International HapMap Consortium used genotyping platforms including Illumina BeadArray to obtain SNP genotype calls from 269 DNA samples across four collections: 30 parent–offspring trios from Ibadan, 30 trios from Utah, 45 unrelated individuals from Beijing and 44 from Tokyo. Parent–offspring trios and the PHASE algorithm supplied haplotype estimation, rather than direct observation of every chromosome. In Figure 7, 36 SNPs resolve seven haplotypes in a chromosome 2 region in the Utah panel; seven tag SNPs capture the 36 SNPs at . The readout demonstrates reusable allelic correlation, not causal effects.
Controls required over 80% complete calls, at most one discordance among five duplicate genotypes, at most one Mendelian segregation error per trio panel, and a Hardy-Weinberg equilibrium test with ; blind reassessment checked performance. Minor allele frequencies of at least 0.05 were targeted, the collections were not population-representative, and inferred haplotypes retain phase uncertainty. These limits restrict extrapolation of linkage disequilibrium.[ref]
A haplotype map of the human genome by The International HapMap Consortium. pp. 1300–1302, Table 1; Figure 7, p. 1306; Methods, p. 1316
Meiosis 2026-10-13
Meiosis reduces two sets of chromosomes to one. In the ordinary diploid life cycle, this makes it possible for fertilization of two haploid gametes to restore the diploid state rather than double the number of sets in every generation. Its two divisions perform different tasks: meiosis I separates homologous chromosomes, whereas meiosis II separates sister chromatids. One round of DNA replication precedes both divisions; there is no second round between them. See [ref]
Molecular Biology of the Cell, 4th edition. Meiosis, “Duplicated Homologous Chromosomes Pair During Meiosis” and “Gametes Are Produced by Two Meiotic Cell Divisions”, Figures 20-6–20-7
.Two homologous chromosomes carry corresponding genetic loci, potentially with different alleles; two sister chromatids originate by DNA replication of one chromosome. After chromosomal crossover between nonsister chromatids, the sister chromatids need not carry identical alleles everywhere. Separating the two relationships explains why a haploid product of meiosis I still contains replicated chromosomes.
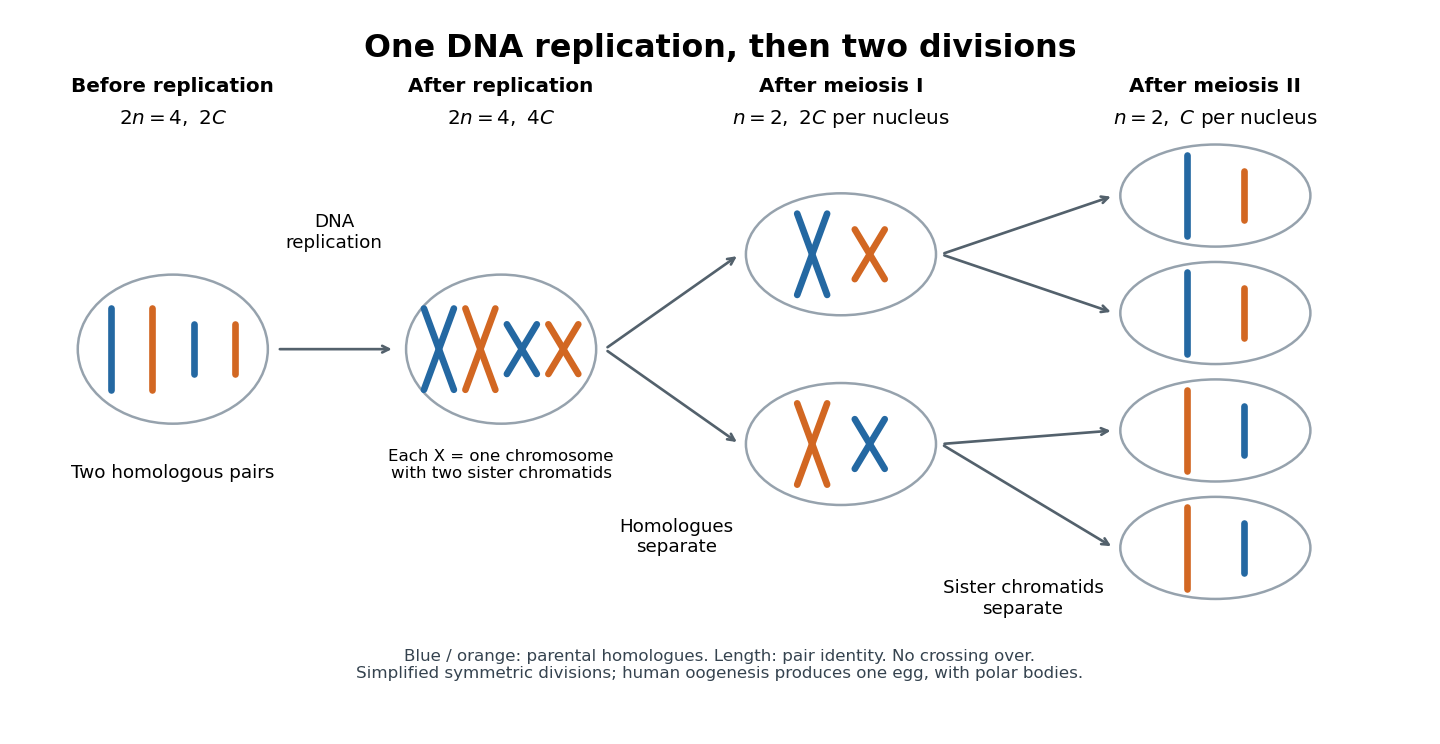
One DNA replication followed by homologous chromosome separation and separation of sister chromatids
. Original simplified model with two homologous chromosome pairs, no chromosomal crossover, and symmetric divisions. The four final compartments represent haploid products, not four functional ova. Human oogenesis uses unequal divisions, retaining most cytoplasm in one developing ovum and producing polar bodies. Details.
The two divisions explain Mendelian segregation; variable orientation of different homologous chromosome pairs connects it to independent assortment. Genetic recombination adds another source of allele combinations. These mechanisms reshuffle inherited material without requiring a new mutation.
The symmetric diagram is useful for counting, but human oogenesis produces one functional ovum rather than four. The polar bodies receive discarded chromosomes and little cytoplasm. See Human Reproductive Anatomy and Gametogenesis (§43.3, Oogenesis and Figure 43.13: unequal divisions and polar bodies).
Meiosis I Created 2026-10-13 Updated 2026-10-20
The first division of meiosis separates replicated homologous chromosomes; their sister chromatids remain associated.
Meiosis II 2026-10-13
The second division of meiosis separates sister chromatids, without an intervening round of DNA replication.
Meiotic drive 2026-10-20
Transmission distortion favouring an element over the Mendelian expectation. In a strict usage this acts during meiosis; broader usage also includes biased survival of gametes.
Mendelian segregation 2026-10-13
In ordinary diploid meiosis, the two copies at a genetic locus separate: each gamete receives one copy. With fair transmission, a heterozygote produces gametes carrying or with probability each. This is a transmission mechanism conditional on a parental genotype, rather than a statement about how common either allele is in a population.
For the one-locus cross , assume independent parental transmissions, no new mutation, and no differences in fertilization or survival between the resulting genotypes. Multiplying the two gamete probabilities gives four equally likely ordered outcomes: , each with probability . Combining the two orders of the heterozygote givesOnly under the additional ideal model of complete dominance of , complete penetrance, and no environmental changes to the scored characteristic do and share the dominant phenotype. Its probability is then . The inheritance calculation alone does not determine a phenotype or predict a generic disease risk.
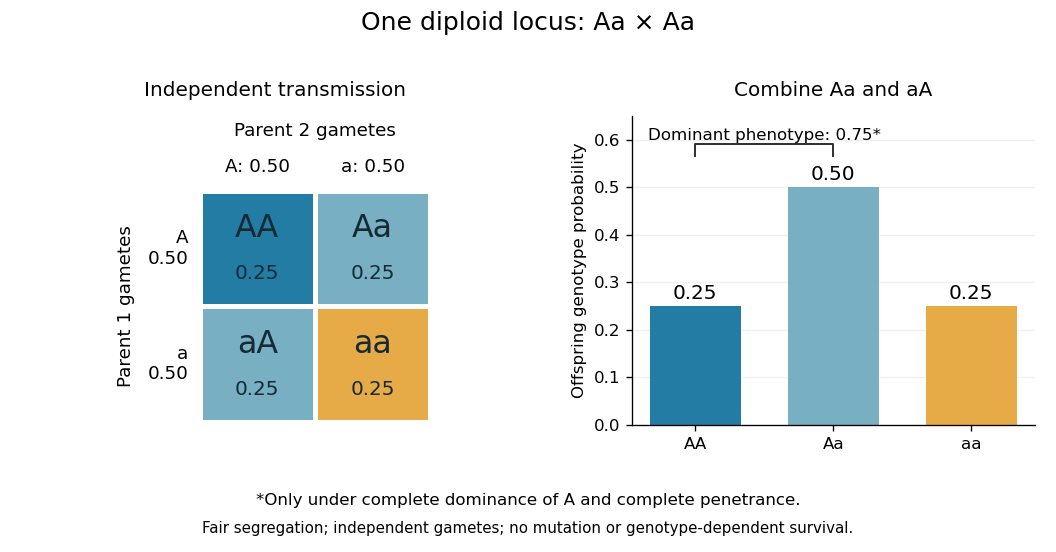
Fair Mendelian transmission in an Aa × Aa cross, with phenotype grouping under complete dominance and penetrance
. Details. 
These are probabilities, not a requirement that every four offspring contain one , two and one . For 100 independent offspring under the ideal model, the expected values of the three counts are ; the count has a binomial distribution with standard deviation . Each new transmission keeps the same conditional probabilities given the parental genotypes.
In his paper published in 1866, Mendel reported 5,474 round and 1,850 wrinkled pea seeds: round, close to the ideal . Following descendants also distinguished two inherited classes within the round phenotype, illustrating why genotype cannot always be read from appearance.[ref] At several genetic loci, transmission patterns additionally depend on genetic linkage and genetic recombination.
Oogenesis 2026-10-13
In ciliates, sex often renews and reorganizes nuclei without immediately increasing cell number, and its outcomes depend on both germline DNA and the old somatic nucleus. nuclear dimorphism is central. A usually diploid micronucleus preserves the germline genome, while a transcriptionally active, often highly polyploid macronucleus controls vegetative cell function. Repeated binary fission can increase cell number without a sexual event; mitotic micronuclear inheritance and often amitotic macronuclear division maintain the two nuclear systems differently. Sex is therefore separable from routine population growth, although some species or strains undergo clonal ageing and benefit from, or eventually require, nuclear renewal for continued vigorous growth.
In the standard conjugation sequence of Tetrahymena thermophila, appropriately mature, compatible cells pair under suitable conditions, often starvation. The micronucleus undergoes meiosis; selected haploid products give stationary and migratory pronuclei after an additional division. Partners exchange migratory pronuclei, and each fuses its incoming and stationary pronucleus to establish a diploid zygotic nucleus. Postzygotic divisions generate new micronuclear and macronuclear lineages, while the old macronucleus is eliminated. The immediate products are two reorganized exconjugant cells, rather than a new single cell created by fusion of the entire parents. Details and product numbers vary across ciliates, so this sequence should not be imposed unchanged on every species.
Compatibility is also more varied than a male/female distinction. Tetrahymena thermophila has seven mating types, with compatibility between different types rather than differentiated sperm and eggs. Mating type is established during formation of the new somatic genome by gene-segment joining and deletion at the mating-type locus; the germline retains genetic potential not all expressed together in the macronucleus. The available outcomes are constrained by the inherited mating-type allele. Primary experiments connecting somatic rearrangements to mating-type expression demonstrate why a simple two-sex Mendelian account is inadequate.[ref]
Selecting One of Several Mating Types through Gene Segment Joining and Deletion in Tetrahymena thermophila by Marcella D. Cervantes and others. Cervantes and colleagues, 2013, results on germline versus somatic mating-type gene organization
Some Paramecium undergo autogamy, in which meiosis and fusion of pronuclei derived from one cell renew the nuclear apparatus without a partner. This can generate pronounced homozygosity rather than introduce new alleles through outcrossing. The potential benefits of recombination and nuclear renewal must therefore be distinguished from the acquisition of genetic diversity from another individual. Sex is not invariably equivalent to outbreeding.
Formation of the new macronucleus also involves DNA elimination, amplification and extensive genome reorganization. Maternal somatic information can influence which sequences survive. In Paramecium tetraurelia, small-RNA-guided excision of the mtA promoter contributes to mating-type inheritance: the old macronuclear state can guide development of the new one.[ref]
Genome-defence small RNAs exapted for epigenetic mating-type inheritance by Deepankar Pratap Singh and others. Singh and colleagues, 2014, abstract and mtA promoter-excision results
This is an experimentally defined epigenetic pathway, not evidence that all acquired traits become heritable. After conjugation, exconjugants can share a newly combined micronuclear genotype yet develop distinct somatic states; later macronuclear assortment can further alter phenotype. Ciliate sex combines meiotic inheritance with somatic-genome reconstruction, and can serve nuclear renewal without immediate numerical reproduction or obligatory outcrossing.Visible divergence and reproductive incompatibility measure different outcomes of evolution. A few changes in a gene regulatory network can transform a conspicuous phenotype, whereas differences in the machinery of meiosis or biological development can prevent reproduction without greatly changing adult appearance. Neither membership of the same genus nor morphological similarity specifies divergence time or the extent of reproductive isolation.
First separate barriers. premating isolation can arise through differences in courtship, habitat or breeding season even when a forced cross produces fertile offspring. postzygotic isolation concerns the survival or fertility of the offspring themselves. Thus fertile captive hybrids do not refute substantial reproductive isolation in nature: they may never encounter suitable mates, may have intermediate ecological requirements, or may be unattractive to both parental biological populations. Under the biological species concept, the relevant issue is effective gene flow among natural biological populations, not the success of one laboratory mating. Conversely, a laboratory failure can reflect husbandry rather than an intrinsic incompatibility.
Rapid visible divergence is especially plausible when sexual selection changes plumage or other signals, or natural selection changes a feeding structure. Such changes can involve a small part of the genome, leaving most interactions required for fertility compatible. In Darwin's finches, genome comparisons associated an ALX1 region with beak shape and also revealed interspecific gene flow. This is an example of conspicuous adaptive differentiation coexisting with exchange of alleles, rather than evidence that all avian hybrids are fertile.[ref]
Evolution of Darwin's finches and their beaks revealed by genome sequencing by Sangeet Lamichhaney and others. Results on interspecific gene flow and the ALX1 haplotype association with beak shape
Continued gene flow can itself help maintain compatibility through much of the genome while divergent natural selection preserves differences at selected genetic loci.For cryptic incompatibility, consider a Dobzhansky-Muller incompatibility. Two lineages start with compatible alleles . One evolves in the presence of , while the other evolves in the presence of . Each change is tolerable in its own genetic background, but the combination disrupts function in hybrids. This requires neither lineage to pass through a low-fertility intermediate. Such epistasis can affect spermatogenesis, chromosome pairing, or interactions between nuclear and mitochondrial genomes, while leaving the adult body plan nearly unchanged. The relevant changes can arise through natural selection, genetic drift, or conflict over inheritance; adaptive divergence in external form is not required.
A particularly informative mammalian experiment concerns PRDM9, which helps specify recombination hotspots. In a particular house-mouse cross, changing its DNA-binding region to the corresponding human sequence restored chromosome pairing and male fertility. The experiment implicates compatibility between a genetic recombination regulator and its binding sites, rather than visible morphological divergence. Crucially, this was a rescue in a specified cross and genetic background, not a demonstration that PRDM9 explains all mammalian speciation.[ref]
Re-engineering the zinc fingers of PRDM9 reverses hybrid sterility in mice by Benjamin Davies and others. Figure 1 and section “Humanizing Prdm9 restores hybrid fertility”; parental-background and engineered-allele comparisons
Comparative work on birds lacking PRDM9 found relatively stable recombination hotspots associated with functional genomic features.[ref]
Stable recombination hotspots in birds by Sonal Singhal and others. Science 350, pp. 928–932; maps inferred from two finch species and comparisons of hotspot conservation
This suggests a possible difference in the routes to incompatibility: turnover of PRDM9 and its targets is unavailable in those lineages. The inference is mechanistic, however; hotspot stability by itself does not demonstrate that avian hybrids will be fertile.Differences in karyotypes provide another mechanism. A balanced chromosomal rearrangement can have little visible effect in either parent yet cause faulty pairing or unbalanced gametes in a heterozygous hybrid. However, a different chromosome count alone does not prove sterility: the arrangement and segregation behavior matter. Differences in genomic imprinting may also disturb placental development or embryonic growth in mammalian crosses, sometimes producing direction-dependent outcomes. Parent-of-origin growth abnormalities and disrupted genomic imprinting in particular rodent hybrids provide evidence that this mechanism can matter.[ref]
Genomic imprinting is disrupted in interspecific Peromyscus hybrids by Paul B. Vrana and others. Nature Genetics 20, pp. 362–365; reciprocal-cross growth abnormalities and allelic expression
These are hypotheses to test in the relevant lineages, not a universal explanation for the bird–mammal comparison.The sex of the offspring matters. Haldane's rule predicts that, when one sex of first-generation hybrids is absent, rare or sterile, it tends to be the heterogametic sex: typically XY males in mammals and ZW females in birds.[ref]
Sex ratio and unisexual sterility in hybrid animals by J. B. S. Haldane. Journal of Genetics 12, pp. 101–109; formulation of the heterogametic-sex pattern
Recessive incompatibilities on a sex chromosome can be exposed when there is no corresponding second copy; rapid evolution of reproductive functions can contribute too. Therefore a report of fertile avian males is insufficient without testing females. The rule predicts a sex bias in a given cross, not that birds as a class have weaker reproductive isolation.To discriminate these mechanisms, compare multiple independently diverged pairs, match their estimated divergence times using phylogenetics, and avoid treating taxonomic rank as a clock. Perform reciprocal crosses with parental within-lineage controls; measure fertilization, embryonic survival, both sexes' fertility, and second-generation and backcross success. A fertile first generation can conceal hybrid breakdown after genetic recombination disrupts coadapted combinations. Pair karyotype analysis and observations of meiosis with genome mapping and carefully controlled rescue experiments. Field measurements of mating and ancestry then determine whether compatibility translates into gene flow. The contrast is explained by different rates and genetic targets of visible divergence and reproductive divergence, with sex, cross direction, ancestry and assay conditions determining the observed outcome.
DNA repair preserves usable inherited information, while genetic recombination can improve the combinations in which that information is transmitted. Their evolution need not have the production or prevention of diversity as an ultimate goal. natural selection favors consequences for survival and reproduction in particular contexts; novelty is neither automatically beneficial nor something an organism must maximize.
mutation creates new sequence variants; genetic recombination usually rearranges variants already present. A chromosomal crossover can produce a new haplotype without changing the allele frequency at either of two genetic loci. Conversely, DNA repair can restore sequence integrity after damage without eliminating the inherited differences between individuals. In particular, a normal alternative allele is not an error merely because it differs from another individual's sequence. These operations address different features of inheritance.
Unrepaired damage can block DNA replication or transcription, break a chromosome, or generate a harmful mutation. DNA repair thus has immediate benefits to a biological cell and can reduce the future burden of deleterious changes in descendants. In multicellular organisms, somatic mutation can compromise tissue function, while germline mutation changes what offspring inherit. The relevant balance includes the biochemical costs of DNA repair, the probability and severity of damage, and the possibility of DNA repair errors. natural selection for increased fidelity can become weak relative to genetic drift when an additional improvement is extremely small; perfect fidelity need not evolve. Some error-prone damage responses can also improve immediate survival even if surviving descendants carry more mutations.
The biochemical relationship is complementary. homologous recombination repairs a DNA double-strand break by using a related intact sequence as a template. In meiosis, programmed breaks can be repaired using a homologous chromosome, sometimes producing chromosomal crossovers and sometimes noncrossover products. Biochemical identification of Spo11 linked a component of the meiotic machinery to programmed break formation, demonstrating that meiotic diversification can begin with a controlled DNA-damage-and-repair process.[ref]
Meiosis-specific DNA double-strand breaks are catalyzed by Spo11, a member of a widely conserved protein family by Scott Keeney and others. Cell 88, pp. 375–384; purification and identification of the protein covalently associated with meiotic breaks
Crossovers also help establish the physical connections needed for reliable segregation in many meiotic systems. This immediate transmission function can favor the machinery independently of any later benefit from a novel haplotype. DNA repair with sister chromatids in another context can preserve sequence without the same exchange between parental homologues.There are also population-level benefits of reshuffling. Suppose two favorable alleles, and , occur in different lineages as and . With no exchange, their lineages compete and either favorable allele can be lost when the other lineage prevails. genetic recombination can create , allowing selection to combine their benefits; it can also separate a favorable allele from linked harmful changes. This is the core of the Fisher-Muller hypothesis and relates to Hill-Robertson interference, the reduction in the efficacy of natural selection caused by associations among selected genetic loci. Recreating low-load genotypes can also slow Muller's ratchet in finite biological populations.
The bookkeeping illustrates why this is not simply increasing all diversity. Let be haplotype frequencies and be linkage disequilibrium. Under the standard random-mating model with recombination fraction and no other evolutionary forces,For and the other two zero, . At , the next frequencies are and . Each favorable allele still has frequency : the process creates both the favorable combination and its unfavorable counterpart. Subsequent natural selection determines their fates. If the original associations are already advantageous, breaking them can instead impose a recombination load.
In controlled yeast experimental evolution, sexual and asexual treatments differed in their rates and genomic routes of adaptation, with the sexual treatment more efficiently separating beneficial from deleterious changes.[ref]
Sex speeds adaptation by altering the dynamics of molecular evolution by Michael J. McDonald and others. Nature 531, pp. 233–236; sexual versus asexual treatments, fitness changes and mutation trajectories
A mechanistic test should control the costs and bottlenecks of the sexual cycle, compare otherwise matched genetic backgrounds, and use reconstructed changes to distinguish genetic linkage from intrinsically different effects of a mutation. Such results establish a benefit under the tested population regime, not an unconditional advantage of sex in every environment. genetic recombination can disrupt favorable epistasis, and the costs of mating are distinct from those of crossover formation.For an explanation of the evolution of the machinery itself, the association with its bearer matters. A recombination modifier must on average increase the representation of its own descendants, perhaps through improved segregation or through association with combinations that selection favors. Merely increasing long-run adaptability of the species is insufficient. Similarly, a mutator allele can spread in an asexual lineage by hitchhiking with a beneficial mutation it helped generate despite producing many harmful ones; exchange can weaken that association. The advantages of fidelity and of exchange consequently depend on genetic linkage, population size, environmental change and the current supply of beneficial variants.
DNA repair can regulate which exchanges are permitted as well. DNA mismatch repair corrects copying errors and can reject exchange between sufficiently diverged sequences. In bacteria, disabling mismatch-repair functions greatly relaxed a barrier to genetic recombination between divergent lineages.[ref]
The barrier to recombination between Escherichia coli and Salmonella typhimurium is disrupted in mismatch-repair mutants by Christiane Rayssiguier and others. Nature 342, pp. 396–401; comparison of mismatch-repair-deficient mutants with the normal recombination barrier
An experiment separating these roles should compare DNA repair of copying errors with exchange between templates of controlled divergence, using a restored DNA repair gene as a rescue. DNA repair and genetic recombination can therefore be parts of the same evolved solution: preserve functional sequence, transmit chromosomes reliably, and reshuffle existing alleles when doing so improves reproductive success.