Ni(II) is . A square-planar complex can keep its strongly antibonding in-plane orbital empty and has a paired, configuration. A tetrahedral arrangement has two unpaired electrons, .
The important change here is the substituent on the imine nitrogen: Me, Et, then tert-butyl. The phenolic ortho-tert-butyl substituent is common to all three ligands. The small N-Me groups permit coplanar NO coordination in . N-tert-butyl crowds that arrangement and favors twisting the two chelate planes toward a distorted tetrahedron in . N-Et gives a closer energetic competition, so accesses both structures. The observed equilibrium reflects ligand-field stabilization, pairing, steric strain and temperature-dependent entropy; it is not a change in nickel oxidation state.
The negative activation volume is consistent with an associative substitution pathway: an incoming donor of the tetradentate ligand approaches square-planar Ni, giving a five-coordinate transition structure/intermediate before a donor of an original chelate leaves. Successive donor substitution and chelate closure give the tetradentate NO complex. Proton transfers convert the incoming phenolic groups to coordinated phenoxides while releasing the original ligands.
With acid/base forms written explicitly, the net ligand exchange is . The connected four-donor ligand benefits from the chelate effect.
The bulky N-tert-butyl groups in hinder precisely the close donor approach needed for that associative route, and the starting tetrahedral geometry must reorganize to accept the planar tetradentate ligand. Consequently exchange can be too slow to observe under the stated conditions. Failure to observe a new product is a kinetic observation; it does not establish that all tetrahedral nickel complexes are inert or that the tetradentate product is thermodynamically impossible.
The natural route is conjugate-base substitution. Hydroxide reversibly removes an ammine proton:The amido donor promotes loss of X, producing a short-lived five-coordinate intermediate. That leaving-group-sensitive loss is the slower stage. Subsequent trapping/proton transfers are fast and no longer depend on which halide left.
Water capture followed by proton transfer gives the hydroxo complex. Competing SCN capture can occur at either end of this ambidentate ligand, producing Co-SCN and Co-NCS. Any initially captured S-bound material can undergo the base-assisted linkage rearrangement considered next. Water is overwhelmingly abundant, so the stable N-bound product can remain a minor overall product. The slow X-dependent and fast X-independent stages fit loss of X followed by chemistry of a common intermediate; a single rate-limiting direct attack by thiocyanate would not explain that separation naturally.
Begin with the same ammine deprotonation and amido-assisted weakening/cleavage of the Co-S bond, but now SCN is already beside the metal. A solvent-caged pair of five-coordinate cobalt amido species and the departing thiocyanate can reorient and recombine through nitrogen before the ligand diffuses into bulk solution. Reprotonation restores the five ammines and gives A.
Thus the essential sequence isWater capture competes to give Co-OH, but rapid internal return of the already associated ligand accounts for the high yield of A. The slower entry into the reactive conjugate-base/dissociation sequence is followed by faster reorientation, capture and proton transfer. This is linkage isomerization, not oxidation or reduction of cobalt.
The near absence of incoming C label means the coordinated thiocyanate usually retains its molecular identity during linkage change. A mechanism that releases it into a fully mixed solution pool and then captures an arbitrary external thiocyanate would predict much more exchange.
Rapid geminate return within a solvent cage explains the result: the original ligand turns and binds through N before an external labeled ligand can intercept the cobalt centre. A continuously bound intramolecular rearrangement is also compatible with low exchange. Therefore the label result supports an intramolecular/cage-local process but does not alone prove that the Co-S bond stays unbroken throughout.
Low-spin Co(III) is substitution-inert. In acid, coordinated ammine deprotonation is suppressed, so the strongly donating amido conjugate base that assists ligand loss is not generated appreciably. Direct aquation/linkage change consequently remains very slow.
The persistence of the starting S-bound complex therefore supports a base-activated kinetic pathway. It is consistent with the rapid rearrangement in base and does not mean that the S-bound form must be the thermodynamically preferred linkage isomer under every condition.
These are inner-sphere electron transfer pathways distinguished by where Cr(II) attacks the coordinated thiocyanate. Cr(II) is substitution-labile and can replace an aqua ligand to form a bridged Co(III)/Cr(II) precursor.
- Remote attack at nitrogen: form . Electron transfer through the bridge gives Co(II)/Cr(III). Cleavage on the now labile Co(II) side leaves the N-bound product . This is the faster channel.
- Adjacent attack at sulfur: Cr binds the same sulfur that is attached to cobalt, giving a shared-S bridge, . Electron transfer and Co-S cleavage leave , the channel.
In both cases the product Co(II) rapidly loses its ammines to give the hexaaqua ion, whereas substitution-inert Cr(III) retains the bridge-end information. The two chromium linkage isomers are therefore a record of two precursor geometries, rather than evidence that one outer-sphere electron-transfer event somehow selects two arbitrary new ligands afterwards.
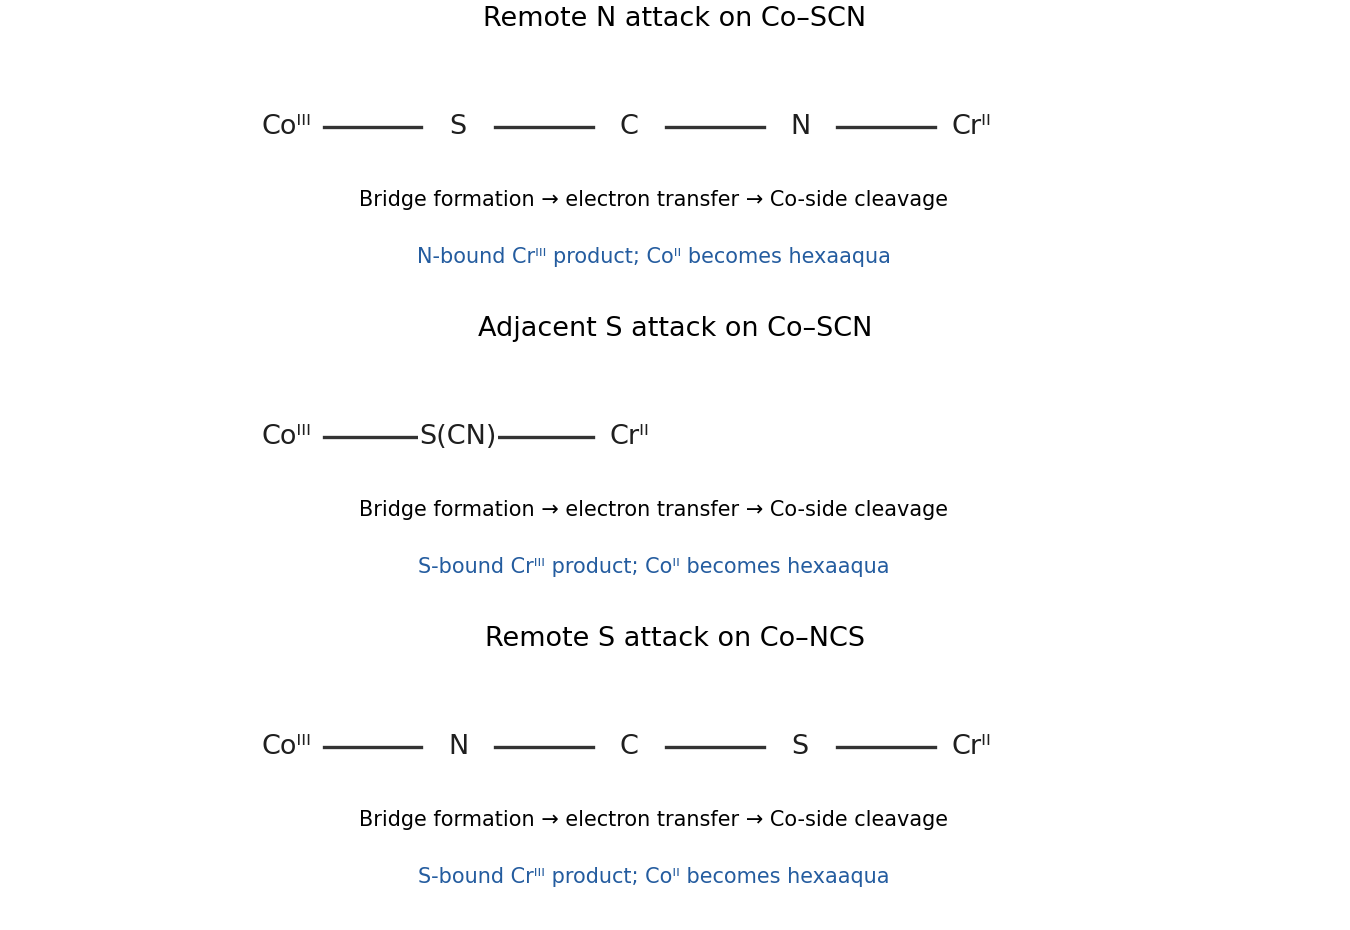
Remote and adjacent bridges retain different thiocyanate linkage information
. Details. The free end of N-bound is sulfur. Cr(II) binds there, giving the remote bridge . Electron transfer and cleavage of the Co-N bond leave S-bound Cr(III), ; labile Co(II) becomes hexaaquacobalt(II).
The much smaller rate constant shows that the efficiency of bridge formation/electron transmission is strongly sensitive to linkage orientation. Efficient donation and coupling in a Co-S-bound precursor need not be reproduced when cobalt is N-bound. Its reduction is still consistent with inner-sphere electron transfer; a slow rate alone is not a criterion for outer-sphere transfer.
Write for the imidazolylidene N-heterocyclic carbene bearing the two mesityl groups shown. The proposed active unit is , with H and SiPh both bonded to nickel, rather than a nickel-free hydrosilane or a bare NHC-Ni fragment.
Displacement of coordinated styrene and formal oxidative addition of Si-H to Ni(0) give a low-coordinate nickel(II) hydride-silyl unit; substrate binding fills a vacant site. The productive cycle is alkene coordination, migratory insertion into Ni-H, then beta-hydride elimination from the other beta carbon, releasing the shifted alkene and regenerating Ni-H.
The strongly donating NHC stabilizes a reactive low-coordinate metal and assists Si-H activation. Its steric bulk discourages unproductive saturated bis-alkene species and promotes product departure, but excessive crowding can hinder substrate binding/insertion. The silyl ligand also tunes the metal electronically: changing its aryl substituents changes donation and the barriers for hydride transfer/elimination. Product release and reversible insertion determine whether further double-bond migration occurs. Bulky ligands can suppress competing alkyl-silyl reductive elimination, which would instead consume substrate by hydrosilylation.
The hydride-silyl formulation and insertion/elimination interpretation are the experimentally proposed mechanism; the individual catalyst resting states need not all be uniquely fixed by the activation recipe.[ref]
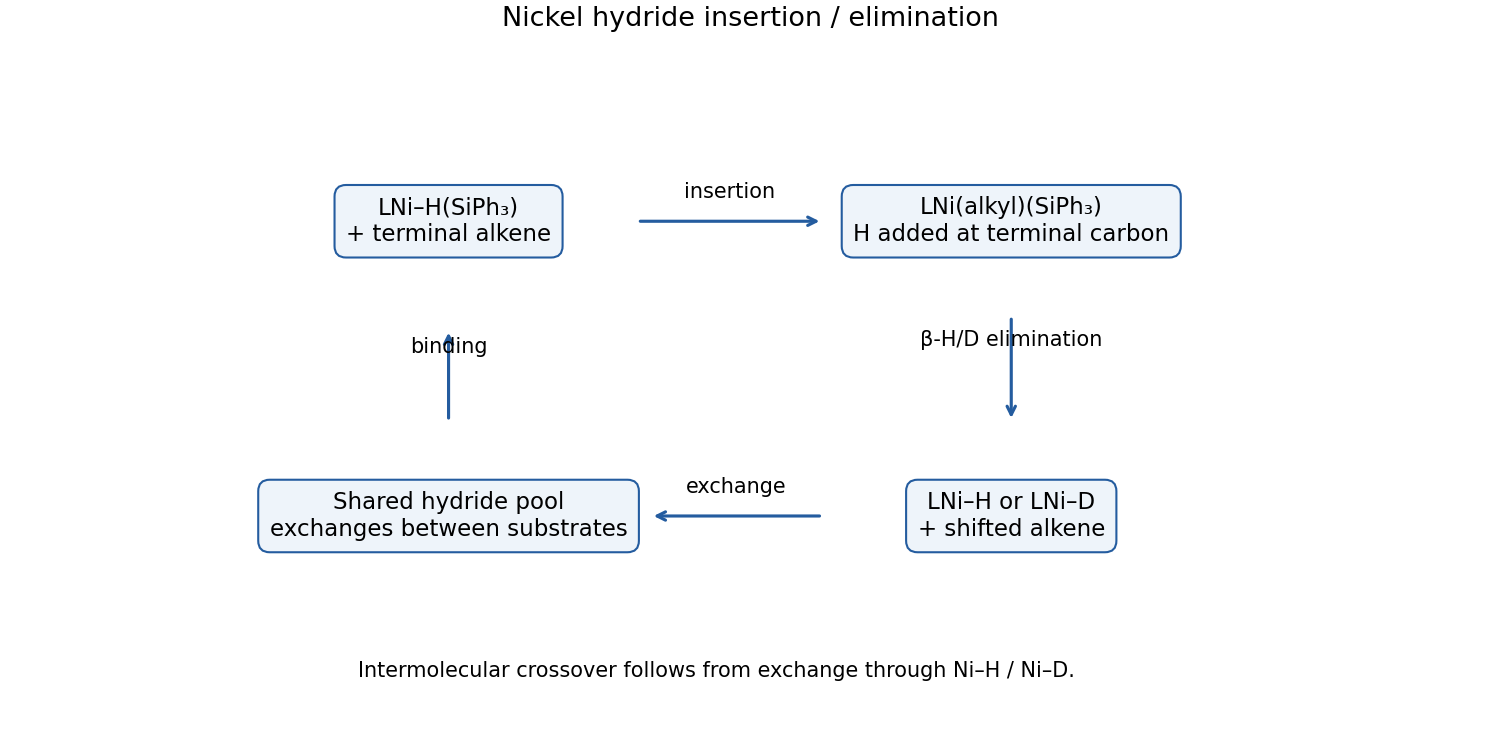
Label the starting exomethylene carbon , the aryl-bearing carbon , the ethyl CH/CD carbon , and the terminal methyl :Insertion into Ni-H places H on and Ni on . Elimination of a beta H or D from gives and Ni-H or Ni-D. Thus the labeled substrate can retain one D on the new vinylic carbon while exporting the other into the catalyst hydride pool.
Ni-D can then add to the exomethylene of either aryl substrate, introducing D into the methyl group derived from . Reversible insertion/elimination and substrate exchange also introduce D into the vinylic position of the originally unlabeled compound. This explains both the approximately comparable methyl labeling of the two products and the larger residual vinylic labeling of the initially CD-bearing substrate.
The decisive observation is intermolecular crossover of deuterium into the unsubstituted-phenyl product. It rules out a purely intramolecular, identity-preserving 1,3-H/D shift. The unequal percentages retain kinetic and substrate-history information; they are not a requirement for completely statistical H/D equilibration and cannot determine all elementary rate constants by themselves.
A Fischer carbene complex typically has a low-oxidation-state, relatively electron-rich middle/late transition metal with strong pi-acceptor ligands such as CO. The carbene is treated as a neutral two-electron donor; metal-to-carbene back-donation contributes to its bond, but its carbon remains electrophilic. An OR or NR substituent stabilizes that carbon by heteroatom donation. Here the metal is Cr(0), , in an 18-electron pentacarbonyl complex.
A Schrock carbene complex usually contains an early metal in a high oxidation state, often , with electron-donating ligands rather than a shell of pi-accepting carbonyls. Its alkylidene has substantial dianionic character in ionic counting, and its carbon is correspondingly nucleophilic. Alkyl substituents are common; heteroatom stabilization of an electrophilic carbon is not the defining feature. These are bonding/electronic limiting descriptions rather than a rule based only on the drawn metal-carbon double bond.
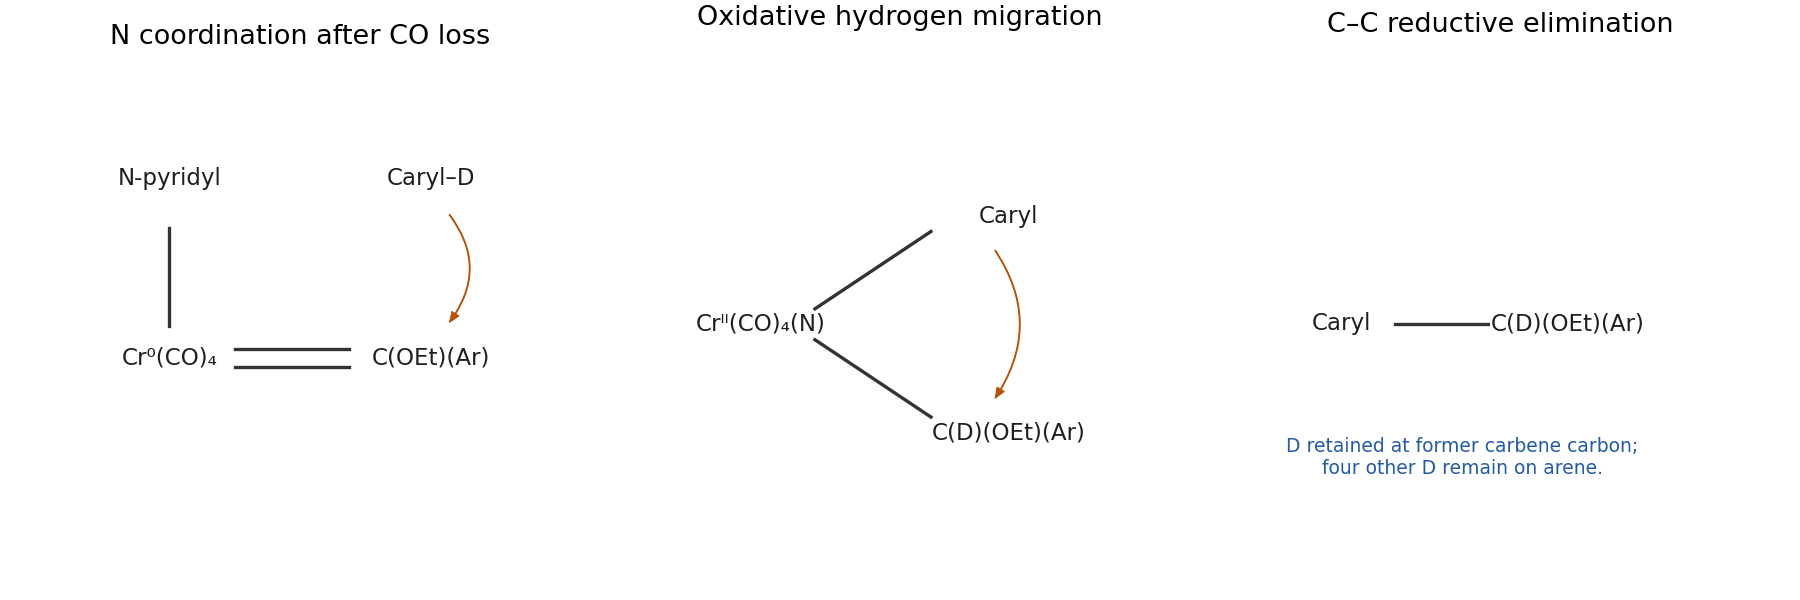
CO first dissociates from the 18-electron Fischer carbene. Pyridyl nitrogen of C then coordinates at the vacant chromium site, positioning an ortho C-D bond of its phenyl group beside the carbene unit.
The appropriate pathway is oxidative hydrogen migration: chromium assists C-D cleavage while D transfers to the electrophilic carbene carbon. The resulting cyclometalated intermediate has both an aryl-Cr bond and an alkyl-Cr bond, the latter to . Formally Cr(0) becomes Cr(II). Reductive elimination between those two carbon ligands forms the new aryl– bond and returns chromium to low oxidation state.
This accounts for all three controls. Extra CO suppresses the coordinative vacancy. Biphenyl E lacks the pyridyl-N directing group and cannot organize the necessary C-H activation geometry. The ortho deuterium moves onto the former carbene carbon rather than being lost to solvent: four D remain on the original phenyl ring and the fifth becomes the benzylic C-D bond.
This is not adequately represented as electrophilic aromatic substitution followed by proton loss, which would discard the diagnostic D. The primary mechanistic study distinguishes oxidative hydrogen migration/reductive elimination from that classical alternative.[ref]
Phosphine binding breaks the hydroxo-bridged dimer into mononuclear Rh-OH units. Hydroxide/boronic-acid association forms a Rh-O-B linkage; transfer of the aryl group from boron to rhodium, with Rh-O bond cleavage, gives the Rh-aryl initiator. This is transmetalation, with rhodium remaining formally Rh(I).
Writing and , the balanced net activation isThe stoichiometric byproduct is boric acid, . Each monomeric Rh(I) product is a 16-electron complex: Rh(I), four electrons from COD, two from phosphine and two from the aryl X ligand. Temporary phosphine dissociation makes room for alkyne binding during propagation.
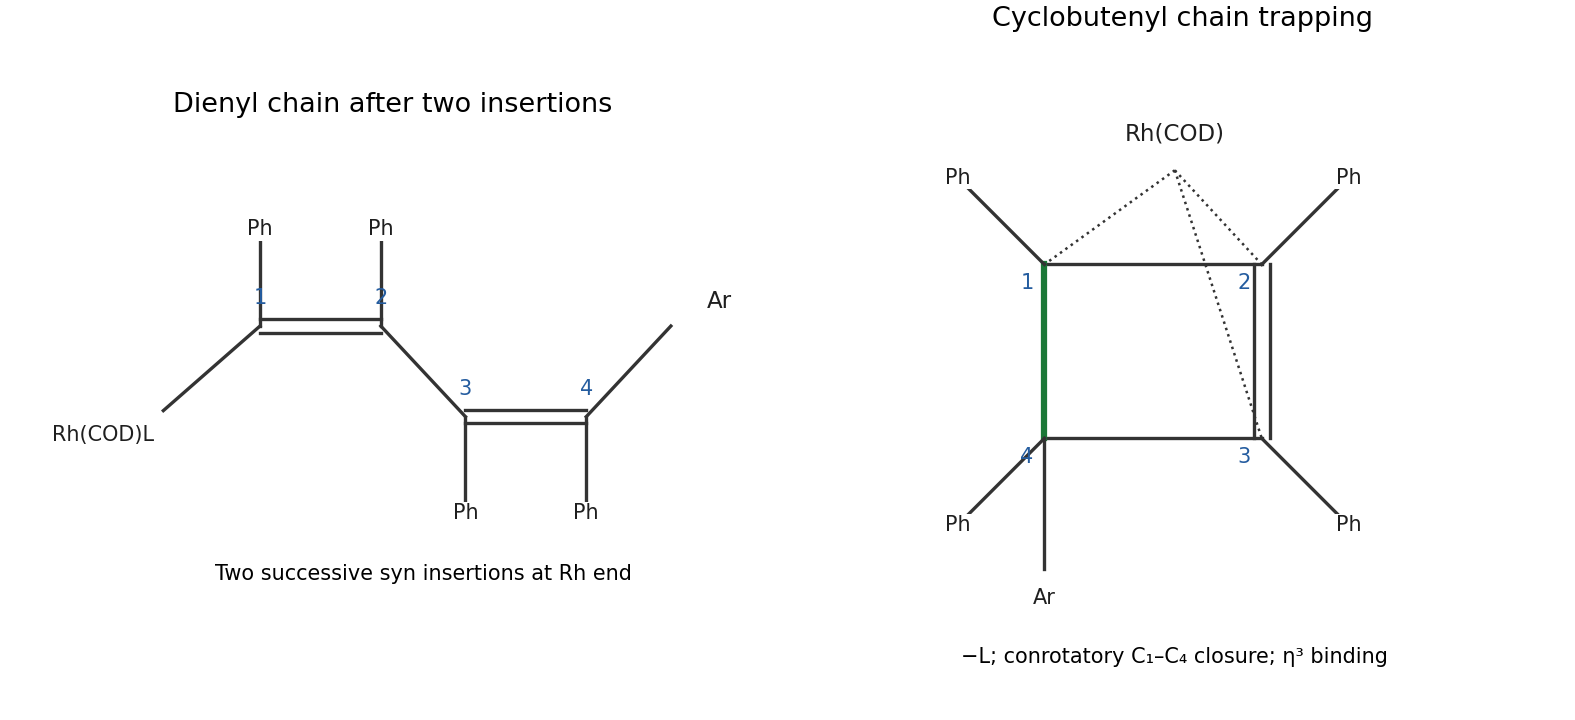
Coordinate one diphenylacetylene and insert it into Rh-Ar. Syn migration puts the new Rh-C and Ar-C bonds on the same face of the former alkyne. Repeat alkyne coordination/insertion into the resulting Rh-vinyl bond. The chain after two insertions isNumbering runs from the metal toward the initiating aryl group. Each insertion is syn: Rh and the continuing chain lie on the same side of the newly made double bond, giving the displayed cis relationship of the two Ph substituents at each inserted unit. The second alkyne inserts at the Rh end; it is not appended by coupling two free vinyl radicals.
To obtain F, promote PPh dissociation, allowing G's dienyl group to fold and form the new C-C bond. A four-pi-electron electrocyclic ring closure gives a cyclobutenyl ligand, stabilized by eta-three coordination of its allylic C-C-C fragment to Rh. The substituted C is the saturated ring carbon bearing Ar and Ph.
The thermal four-pi closure is conrotatory. Ring closure plus eta-three binding restores a 16-electron Rh(I) centre after phosphine loss; no net oxidation-state change is needed. F is an eta-three cyclobutenyl complex, not an eta-four cyclobutadiene with five substituents squeezed onto four unsaturated carbons.
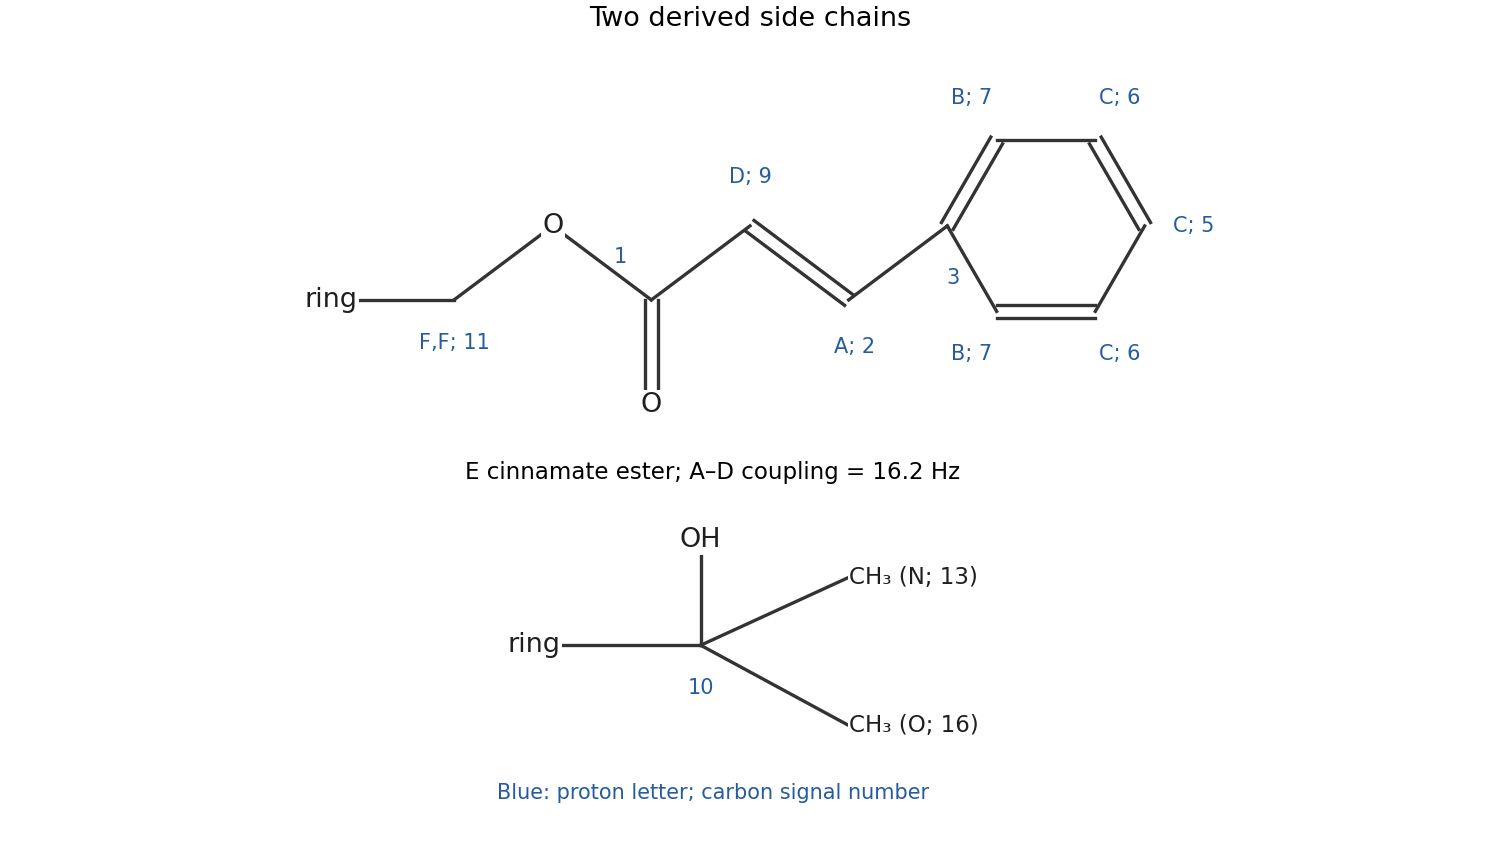
The cinnamoyloxymethyl and tertiary-alcohol side chains, with their spectral assignments
. Details. The two substituents are , with the alkene E, and . The free bond at the left of each drawing attaches to the cyclohexene.
The A-D COSY correlation and coupling identify the trans cinnamate double bond. Five aromatic protons and four phenyl-carbon environments identify its monosubstituted phenyl ring. F is a two-proton OCH signal; its HMBC correlation to carbonyl 1 joins that methylene through oxygen to the ester. The two three-proton singlets N/O, their correlations to oxygenated quaternary carbon 10, and the missing OH resonance give the tertiary-alcohol substituent.
On the ester-containing side chain, read outward from the cyclohexene:A is the downfield beta-vinylic proton and D the alpha-vinylic proton next to the ester.
The phenyl ortho pair is B on carbon 7; the meta pair is C on carbon 6; its para proton is the remaining C on carbon 5. Thus the C envelope integrates to three protons, while B integrates to two. Carbon 3 is the unprotonated ipso site.
In the other substituent, 10 is the quaternary C-OH carbon. The methyl assignments are N on 13 and O on 16, reading the very closely spaced HSQC cross peaks together with the positive methyl responses in DEPT-135. Their mutual long-range correlations and correlations to 10 support the gem-dimethyl group. The two methyl carbons are distinguishable in this chiral molecular environment; their exact stereotopic names are not determined here.
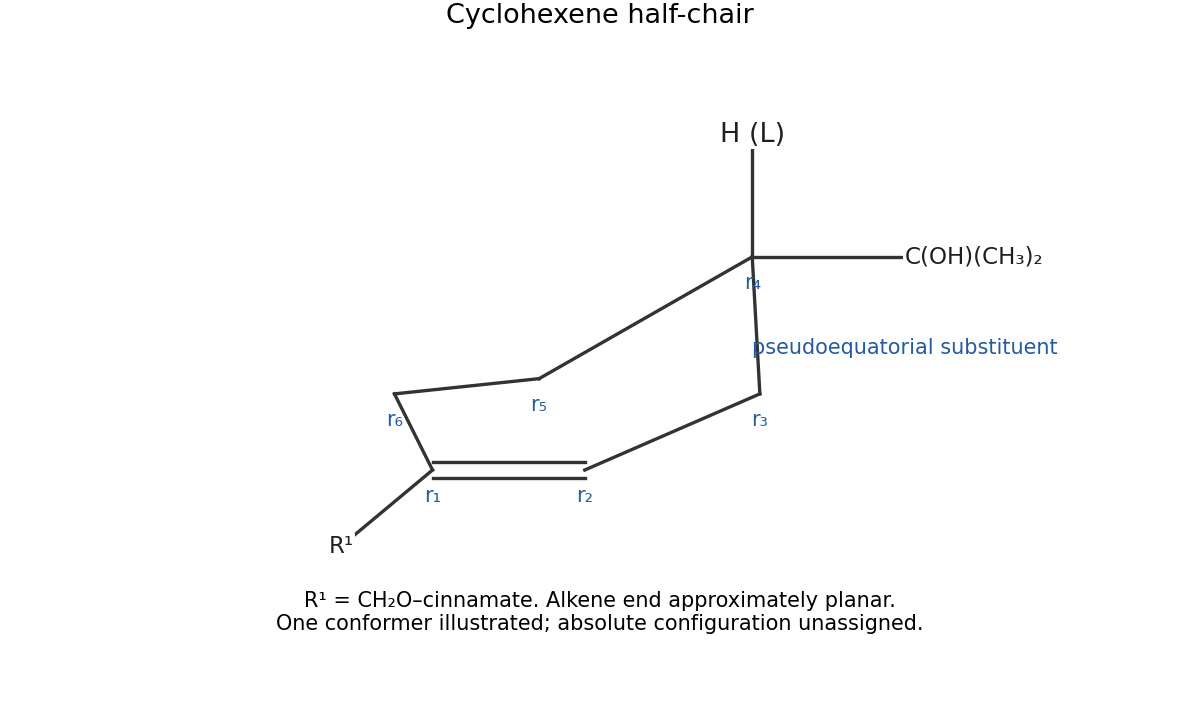


Use a cyclohexene half-chair, not a fully puckered saturated-cyclohexane chair. The alkene carbons and their directly attached bonds form the approximately planar end; the saturated part puckers. Put the substituent pseudoequatorial at the opposite saturated methine to avoid a crowded pseudoaxial arrangement. Its methine proton is correspondingly pseudoaxial. This is a preferred conformation; the achiral spectra do not assign an absolute configuration.
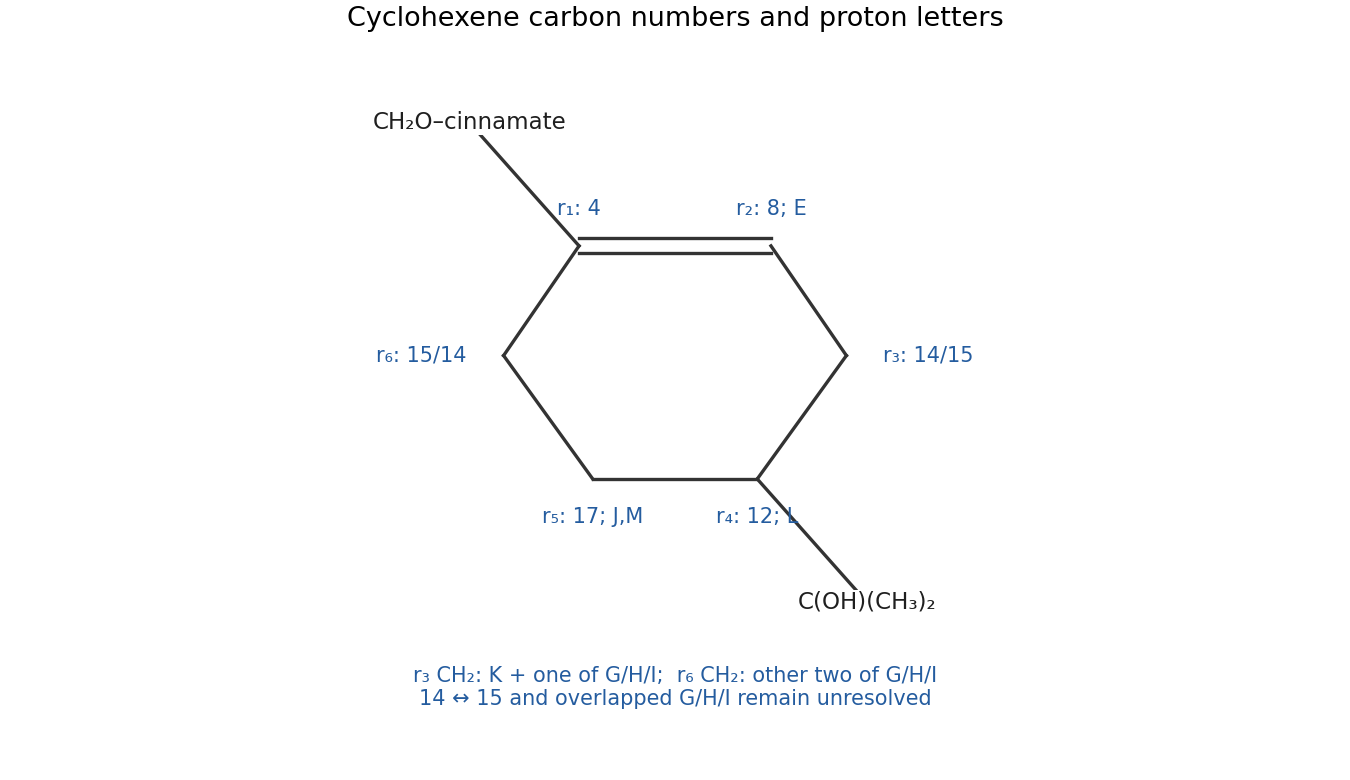
Cyclohexene assignments with explicit ambiguity for the overlapping allylic methylenes
. Details. 
Number ring positions through around the ring, with the double bond. These auxiliary ring-position labels are distinct from the printed carbon-signal numbers.
- : quaternary alkene carbon 4, attached to the OCH side chain.
- : alkene CH 8, proton E.
- : allylic CH, one of carbon signals 14/15; its protons are K and one of G/H/I.
- : methine 12, proton L, attached to the tertiary-alcohol carbon 10.
- : CH 17, protons J and M.
- : the other allylic CH 15/14, carrying the remaining two of G/H/I.
The HSQC pairs J and M with 17 and places L on 12. It also places K and one unresolved member of G/H/I on one of the closely spaced allylic carbons, with the other two G/H/I protons on the other. COSY connects the vinylic E to the adjacent allylic part and connects the methine L to its neighboring methylenes; HMBC links F to the substituted alkene end and the gem-dimethyl group to the methine end.
The printed overlap does not justify a unique assignment of every G/H/I letter or a forced 14-versus-15 assignment to those two ring positions. The drawing retains those alternatives explicitly. In the half-chair, L is pseudoaxial; the resolved large/small couplings are consistent with pseudoaxial/pseudoequatorial methylene partners, but exact axial letter labels within the overlapped group remain uncertain.
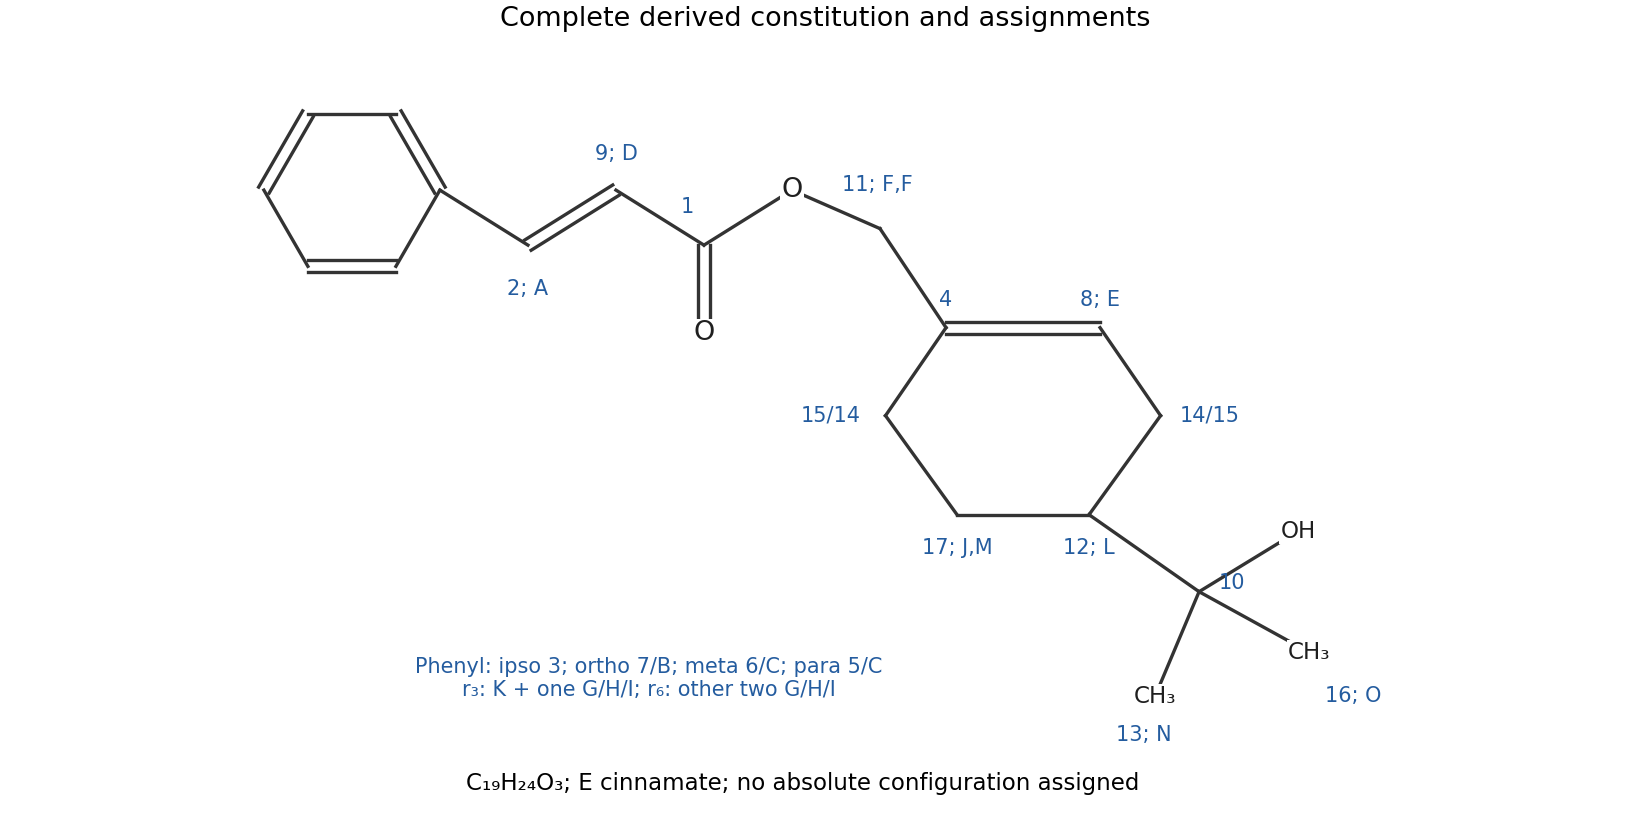
Complete constitution: cinnamoyloxymethyl at cyclohexene position 1 and a tertiary-alcohol substituent at position 4
. Details. 
The constitution is [4-(2-hydroxypropan-2-yl)cyclohex-1-en-1-yl]methyl (E)-3-phenylprop-2-enoate, with absolute configuration at the saturated substituted ring carbon unspecified.
The OCH side chain is attached to the unprotonated alkene carbon 4: F correlates through the ring junction to the alkene carbons and an allylic methylene, while its correlation to 1 fixes the ester linkage. The side chain is attached to methine carbon 12, as shown by the methyl-to-12 HMBC correlations and the contiguous ring COSY network.
The formula check gives 19 carbons and 24 hydrogens including OH. Its degree of unsaturation is eight: four from phenyl, one ester carbonyl, one cinnamate C=C, one cyclohexene C=C and one ring. No additional double bond or hidden second carbonyl is needed.
A warmer troposphere does not imply a warmer stratosphere: increased carbon dioxide can cool the middle and upper stratosphere by enhancing infrared emission. One direct kinetic effect is slower temperature-dependent destruction in the Chapman cycle, tending to increase upper-stratospheric ozone. Two indirect chemical effects can have the opposite sign. If the entry atmospheric cold trap warms, it can admit more water; reaction with an electronically excited oxygen atom then supplies more of the HOx family, enhancing catalytic ozone loss. In cold polar air, changes in water and temperature can favour polar stratospheric clouds, whose surfaces activate halogen reservoirs and can enhance ozone destruction when sunlight returns. These are conditional pathways, not a universal sign for total ozone: cold-trap temperature and circulation, halogen abundance and altitude all matter. Methane oxidation can also supply stratospheric water.[ref]
Scientific Assessment of Ozone Depletion 2022 executive summary. Executive summary: Stratospheric Temperature, water-vapour influences, and temperature-dependent polar ozone loss
For one specified channel, the photolysis rate coefficient isIt depends on spectral actinic flux, absorption cross section, and channel quantum yield. Solar zenith angle, season, overhead absorbers, clouds and aerosol scattering affect the photon field; branching and collisional quenching affect the chemical yield. Absorption alone does not guarantee the desired dissociation.
Curve A is oxygen photodissociation; curve B is ozone photodissociation. Molecular oxygen needs relatively short-wavelength ultraviolet photons, which are strongly screened at lower altitudes, so its rate increases steeply with height. Ozone can dissociate using longer wavelengths that penetrate much further, producing the much larger, less height-sensitive rate shown by B. The plotted quantity is a first-order coefficient with units , rather than a bimolecular rate constant.
The dominant OH-to-HO pseudo-first-order rate is , corresponding to . Recycling HO through atomic oxygen and NO has rate , corresponding to for an individual HO radical. A fixed-total two-state exchange relaxes at the sum of forward and reverse rates, giving about for the partition perturbation. Its fast partition is thereforeThis supports a steady-state approximation in chemical kinetics for rapidly recycled intermediates if the parent concentrations vary slowly. It does not by itself establish stationarity of the entire HOx family. The later source–termination estimate gives a family turnover time of about 16 hours, far longer than partition relaxation; a stationary absolute abundance additionally assumes sufficiently persistent radiation and background conditions. This is the distinction between fast family partition and slow family balance.
Let denote the rate of reaction . For odd oxygen defined as O plus O, reactions 5 and 6 each consume one unit, while NO photodissociation in reaction 7 restores one. Thus the loss from reactions 4–8 is . HO stationarity gives , and NO stationarity within this restricted recycling loop gives . ThereforeThe requested expression neglects the small termination correction in the odd-oxygen budget, while retaining it to determine the much slower total-radical balance. This is consistent when cycling greatly exceeds termination. The net main catalytic cycle is O plus O forming two O molecules; the NO-assisted route mainly interconverts O and O and cancels from this restricted net loss after NO recycling.
Using an isothermal atmosphere with the stated scale height gives and at . The water atmospheric mixing ratio then gives . Applying steady state to the short-lived electronically excited oxygen atom,Its HOx family source is . Balancing this with termination gives . Combining with the rapid partition, or solving both stationary radical equations including termination, givesConsequently the main catalytic odd-oxygen loss isThe Chapman cycle loss is , so the given HO cycle is about 2.35 times larger. Both rates count two odd-oxygen units per completed O-plus-O destruction event. The omitted correction is only about of the main HO loss. The family abundance divided by its source is about , explaining why its absolute stationary state is a stronger assumption than rapid OH/HO interconversion.
With , a reaction of an electronically excited oxygen atom with hydrogen gives one OH and, after rapid H conversion, one HO. The radical source becomesThis is about 4.75 times the original source. Holding water, oxygen, NO and radiation fixed keeps the fast radical partition approximately unchanged, while bimolecular termination makes total radical abundance scale as . Thusabout 2.18 times the original catalytic loss. This predicts a stronger ozone-loss tendency within the specified stationary box model, not a numerically determined percentage change in the ozone layer. The latter requires coupled odd-oxygen production, changing ozone concentrations, transport and the longer-term water budget; those have been held fixed here.
Peroxyacetyl nitrate forms reversibly from a peroxyacetyl radical and NO. It stores reactive nitrogen while intact, then thermal dissociation can release it again. It can therefore transport nitrogen through colder free-tropospheric air and recycle it more rapidly in a warmer planetary boundary layer. Deposition or further chemistry can compete, but reversible storage is its reservoir role.
Dinitrogen pentoxide is likewise a reservoir because can return reactive species. Conversion to nitric acid makes it a sink from the active nitrogen oxides family, especially through heterogeneous uptake on wet aerosol or cloud particles, followed by deposition or washout. Water vapour alone and wet-particle uptake are different mechanisms; they should not be assigned the same rate constant. Temperature, particle surface area and removal are often very different between the boundary layer and the free troposphere, so a reservoir can be transported in one environment and chemically removed in another.
For the stated total number density, multiply the number atmospheric mixing ratio by . With NO defined as NO plus NO,The water partial pressure is of the total pressure. It is a 3% number fraction, not 30 ppm.
The fast family is NO plus NO. Put , , and freeze the supplied parent concentrations while definingThe two equations are and . Their stationary solution isHere , , and in inverse seconds; . Reversible storage must not be counted as permanent family loss: only the term represents the association route followed by hydrolysis.
Closed-family interconversion has . Including the supplied losses, the two linear relaxation times are approximately and , from the eigenvalues of . Fast exchange is therefore well separated from the slower total-family adjustment. However, NO is consumed by ozone on , comparable to and shorter than the family relaxation. Thus the quoted stationary concentration is justified for a maintained background or a sufficiently slowly varying quasi-stationary environment; it is not automatically reached in an isolated parcel initialized with the supplied composition. NO auto-oxidation is negligible at this concentration. Additional organic losses of NO, if present, would need to be added to the model.
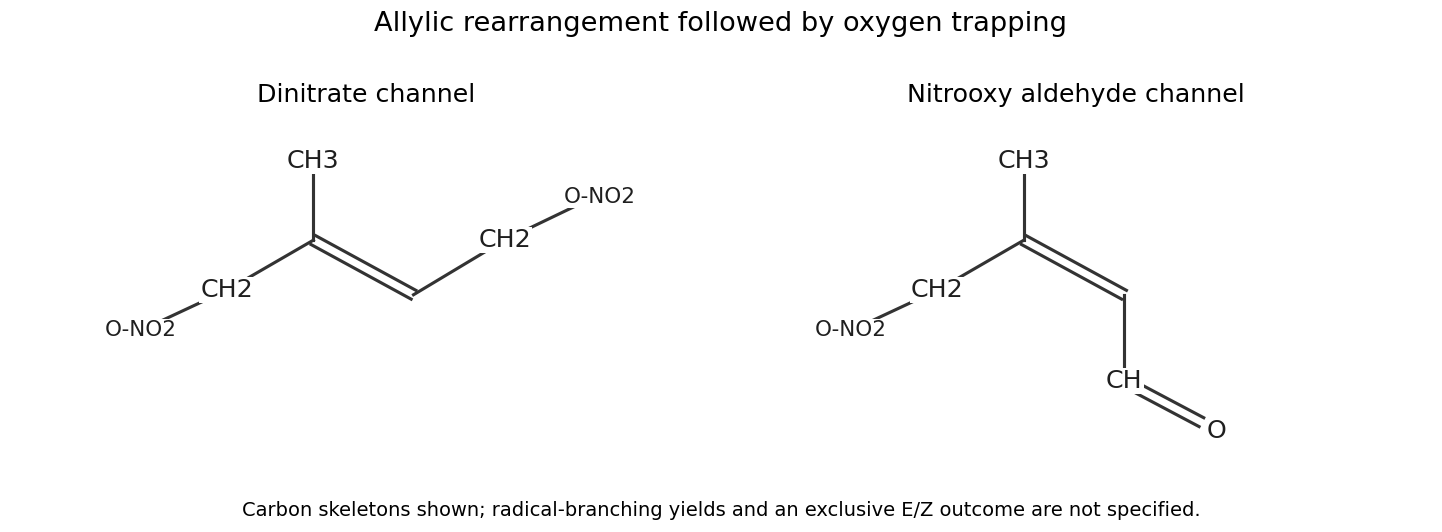
At night, nitrate radical addition and ozone attack are relevant oxidations of isoprene; direct daytime photochemistry is absent, though small OH sources can still exist. The nitrate-containing products identify the NO-addition route. Add NO to the terminal carbon next to the methyl-substituted alkene carbon, producing a nitrooxy substituent and an allylic radical. Resonance moves the radical to the far terminal carbon while shifting the double bond. Oxygen trapping gives a peroxy radical with the carbon skeletonA nitrate-forming termination branch, for example the organic-nitrate branch of reaction with NO, replaces the terminal peroxyl group by a nitrate ester, giving . An alternative branch forms an alkoxy radical, such as peroxy-radical reaction with NO producing RO, NO and O. Reaction of the terminal CHO group with O then gives an aldehyde plus HO, producing . These are covalent O–NO ester groups, not C–NO nitro groups. The delocalized radical can be trapped after conformational change, so this mechanism does not require one universal alkene geometry for both product channels. Product structures alone do not fix a unique branching ratio.

Activate the oxime hydroxyl oxygen with thionyl chloride to give a good N–O leaving group. In the Beckmann rearrangement branch, the C–C bond anti to that leaving group migrates onto nitrogen as the N–O bond breaks. Here the migrating carbon is the spiro centre. Inserting N between that centre and the former oxime carbon expands the five-membered oxime ring to a six-membered nitrilium ion ring. Water attacks the electrophilic nitrilium carbon, followed by proton transfers and tautomerization to the observed spiro lactam. The lactam carbonyl oxygen is introduced in the aqueous work-up.
The competing Beckmann fragmentation cleaves the same anti C–C bond instead of completing migration onto N. Formation of the C≡N bond gives an open-chain nitrile and a tertiary carbocation at the former spiro centre. Loss of a proton from the adjacent chain carbon then gives the exocyclic alkene attached to the retained cyclopentane. The key distinction is N insertion and ring expansion for the lactam, versus C–C cleavage followed by elimination for the unsaturated nitrile. Stabilization of the resulting tertiary carbocation makes fragmentation plausible.
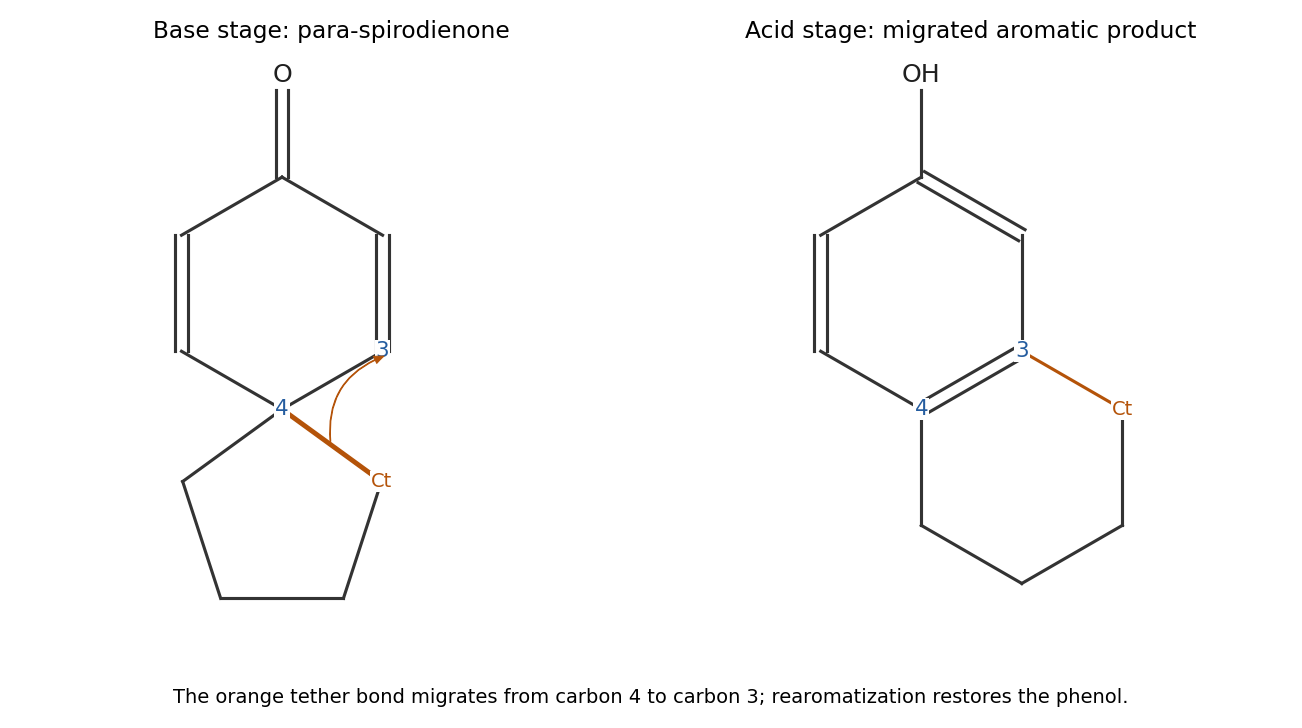
Base first gives phenoxide. Use its para-carbanion/quinoid resonance contributor: the para carbon bearing the tether attacks the terminal chloromethylene carbon in an intramolecular substitution, ejecting chloride. This forms a five-membered ring at that carbon and produces a para-spirocyclohexadienone, with the phenoxide oxygen becoming a carbonyl oxygen. The first stage is a dearomatizing spirocyclization of a phenoxide.
On acidification, carbonyl protonation makes the dienone electrophilic. A tether C–C bond attached to the spiro centre migrates to the adjacent dienone carbon, with the accompanying pi-bond shifts; loss of a proton restores aromaticity. This dienone-phenol rearrangement converts the spiro five-membered ring into the fused six-membered ring while regenerating the phenolic OH. The apparent bond formation meta to OH is thus explained by para spirocyclization followed by skeletal migration, rather than requiring the final fused connectivity to form directly in the base step.

Amide base removes a proton alpha to the nitrile, giving a resonance-stabilized carbon nucleophile. Removal of the available aryl proton ortho to chlorine, together with chloride elimination, generates a benzyne between the chlorine-bearing carbon and its unsubstituted ortho neighbour. The side-chain nitrile carbanion attacks the former chlorine-bearing aryne carbon intramolecularly; one aryne pi bond shifts to the neighbouring aryl carbon. Protonation of that aryl anion by ammonia restores the aromatic ring. The newly formed C–C bond closes the four-membered ring of the cyanobenzocyclobutane product. Two deprotonation processes are required to generate both the nitrile nucleophile and the aryne. The nitrile group is retained, and the product is not formed by simple replacement of aryl chlorine with an external amide nucleophile.
Articles were limited to the first 100 out of 201 total.
Articles by others on the same topic
There are currently no matching articles.