The product is 1,4-dimethoxy-2,5-bis(2-methylbutan-2-yl)benzene. Each new substituent is , a tert-amyl group, rather than the unrearranged secondary alkyl group of the alcohol. The two substituents are related by the symmetry of the ring.
The proton assignments are:
| chemical shift / ppm | Assignment | Reason |
|---|---|---|
| Two aromatic H, at positions 3 and 6 | symmetry makes them equivalent; neither has an adjacent aromatic H to give a resolved ordinary ortho splitting. | |
| Two methoxy groups, six H | Equivalent groups give a NMR singlet. | |
| Two ethyl groups, four H | Each couples to a terminal three-proton methyl group, giving a NMR quartet. | |
| Four methyl groups attached to the two quaternary benzylic carbons, twelve H | These methyl groups have no neighboring methine group proton and give a NMR singlet. | |
| Two terminal ethyl methyl groups, six H | Each couples to two methylene group protons, giving a NMR triplet with the same coupling constant as the NMR quartet. |
The integrals add to 30 H. The unusually shielded terminal ethyl methyl signal is compatible with an aryl-containing environment; its NMR quartet–NMR triplet partner and integral are the decisive assignment evidence. The molecular formula and degree of unsaturation, , fit a benzene ring with saturated side chains.
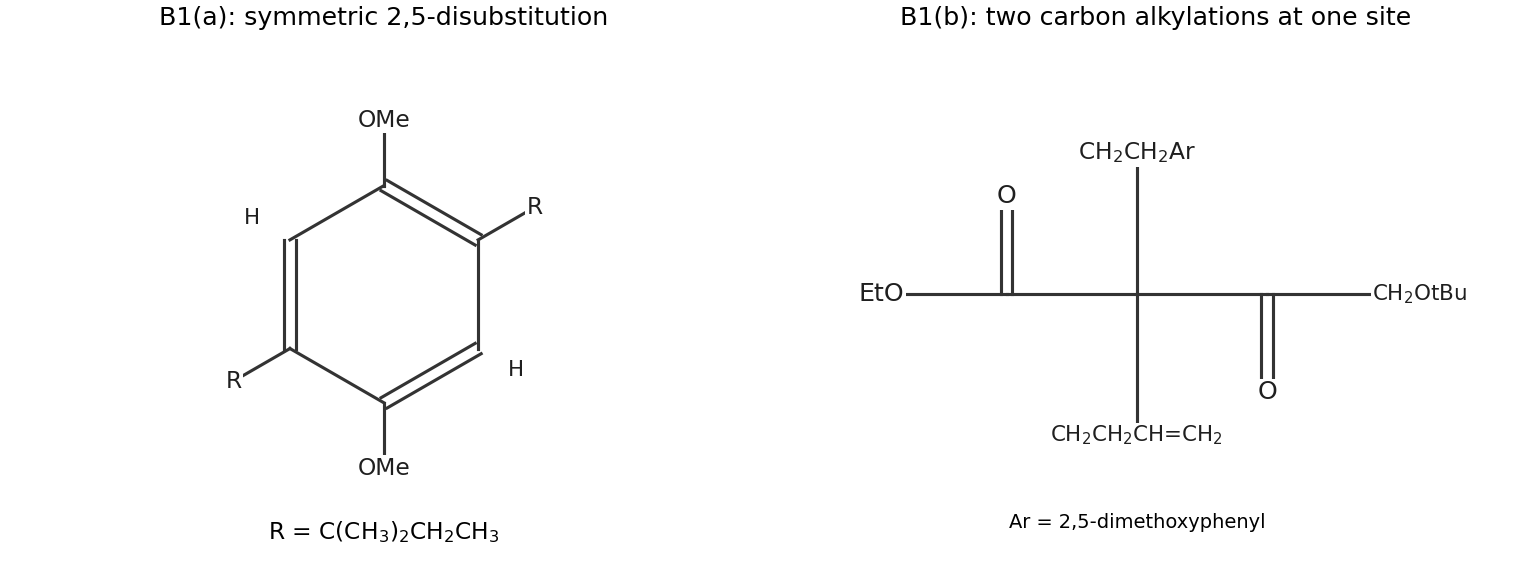
Original structures of the rearranged aromatic alkylation product and dialkylated beta-ketoester
. Details. protonation of the alcohol converts its OH group into a water leaving group. Departure of water initially gives the secondary carbocation . A 1,2-hydride shift from the neighboring methine group transfers its C–H bonding pair to the electron-deficient carbon:The resulting tertiary carbocation is the tert-amyl electrophile. An aromatic bond of the dimethoxybenzene attacks that carbon, producing a Wheland intermediate. Removal of the proton from the attacked ring carbon restores aromaticity and regenerates acid. Repeating this Friedel-Crafts alkylation at the symmetry-related second site gives the disubstituted product.
The hydride migration explains the new ethyl group and the absence of a benzylic methine group proton. Simply attaching the original alcohol skeleton at its OH-bearing carbon would not explain the twelve-proton NMR singlet.
The methoxy groups activate the aromatic ring by electron donation through chemical resonance and direct electrophilic substitution ortho and para. Their para positions are already occupied by one another, so the first alkylation occurs at an available ortho position; the four initially vacant positions are equivalent by symmetry.
After placing the first tert-amyl group at position 2, position 5 is favored for the second substitution. It is para to that bulky group and ortho to the other methoxy group. Position 3 is crowded between methoxy and tert-amyl substituents, while position 6 is near the first tert-amyl group across the intervening methoxy-bearing carbon. The 2,5 arrangement distributes the bulky groups on opposite parts of the ring and preserves the equivalence seen in the spectrum.
Alkyl groups do not electronically deactivate the ring, so the stopping point needs a steric explanation. After two such substitutions the remaining positions are much more congested, each adjacent to a bulky tert-amyl group and a methoxy group. Two large substituents are accommodated much more readily than a third, despite excess alcohol. The formula and NMR identify this sterically favored disubstitution; they do not imply that a Friedel-Crafts reaction always stops after two alkylations.
Let denote the 2,5-dimethoxyphenyl group. A is the doubly C-alkylated beta-ketoesterThe central carbon has no hydrogen after the second step. The alkene, ester and tert-butyl ether remain intact.
carbonate reversibly removes a proton from the methylene group between the two carbonyl groups. The resulting resonance-stabilized enolate attacks the primary iodoethyl carbon of the aryl-containing reagent by an displacement, releasing iodide. The monoalkylated intermediate still has one proton at the same activated carbon. A second deprotonation followed by attack on the primary bromide introduces the but-3-en-1-yl group.
The beta-dicarbonyl position is much more acidic than the methylene group next to the ketone and ether at the other end, explaining the alkylation site. Counting the retained carbon skeleton gives , including the two carbonyl groups and one terminal alkene. The second panel of the original product drawing specifies this connectivity.
The new ring is formed by intramolecular Friedel-Crafts addition to the ketone. titanium tetrachloride coordinates the ketone oxygen of the cyclic beta-ketoester, increasing electrophilicity at that carbonyl carbon. The nearby electron-rich aryl ring attacks this activated ketone at the available position ortho to its ethyl tether.
The electron-flow sequence is: an aromatic bond forms the new aryl–carbonyl-carbon bond, the ketone bond moves onto oxygen, and loss of the aromatic proton restores aromaticity. The two-carbon tether and the shared quaternary carbon make the newly closed carbocycle six-membered. Hydrolytic work-up converts the titanium-bound alkoxide into the tertiary alcohol.
The lactone carbonyl survives. This is an addition to the ketone with retention of its oxygen as OH, rather than an acyl substitution that removes that oxygen. The pendant terminal alkene is not involved. These bond changes account for the fused aromatic ring system and the alcohol in C without needing to assign the new stereochemistry.
Number the enone carbonyl carbon C1, the adjacent alkene carbon C2 and the ethoxy-bearing alkene carbon C3. In the bromine-first pathway, the enol-ether-like unit attacks at C2, breaking the Br–Br bond. The intermediate has a new C2–Br bond and positive charge at C3, stabilized as an oxonium chemical resonance contributor by donation from the ethoxy oxygen.
triethylamine removes the remaining C2 hydrogen. Its C–H bonding pair reforms C2=C3 and neutralizes the oxonium system. The result is replacement of the vinylic C2–H by C2–Br, with the original enone and ethoxy group retained. If bromide first captures the cationic intermediate, the equivalent vicinal dibromide can eliminate HBr to reach the same product.
The ethoxy group makes this alkene sufficiently electron-rich for the electrophilic-addition description. It is important to place bromine at C2 and restore the double bond, rather than stop at a saturated addition product.
For an amine-first pathway, use triethylamine as a nucleophile in a reversible conjugate addition. Its lone pair attacks C3 of the enone; electron shifts through C2 and the carbonyl give a zwitterionic enolate with a C3– bond.
The C2 nucleophilic site of that enolate attacks , installing C2–Br and releasing bromide. Removal of C2–H by base then reforms C2=C3 while the C3–N bond breaks back to neutral triethylamine. This addition–bromination–elimination sequence gives the same C2-brominated enone and regenerates the nucleophilic amine, with another base equivalent accepting the eliminated proton.
These are alternative formal sequences requested by the question, not a kinetic determination that one must dominate under unspecified conditions. In particular, the amine-first sequence does not require triethylamine to generate a high concentration of a vinylic carbanion by direct deprotonation.
A is a fused five-/six-membered bicyclic diketone containing sulfur. To specify its structure unambiguously, retain the cyclohexanone numbering C1=O, C2 and C3 for the original alkene. Add the five-membered ringThe original C1 ketone remains, and the former methyl-ester carbonyl becomes the second ketone. The product is drawn in the first panel below.
methoxide first deprotonates the thiol. The soft thiolate attacks the beta carbon C3 in a Michael addition, shifting the enone electrons onto oxygen. The resulting enolate is nucleophilic at C2. It then attacks its tethered ester carbonyl intramolecularly, forming a five-membered ring through a tetrahedral intermediate. Collapse expels methoxide. proton transfers and work-up give the neutral bicyclic beta-diketone.
The initial 1,4 addition followed by intramolecular Claisen acylation explains the regiochemistry: it connects C3 to sulfur and C2 to the ester carbonyl, giving a favorable five-membered closure. Addition to the original ketone would not give this same enolate-and-ester arrangement. The net loss of methanol from the combined starting materials gives the observed formula.
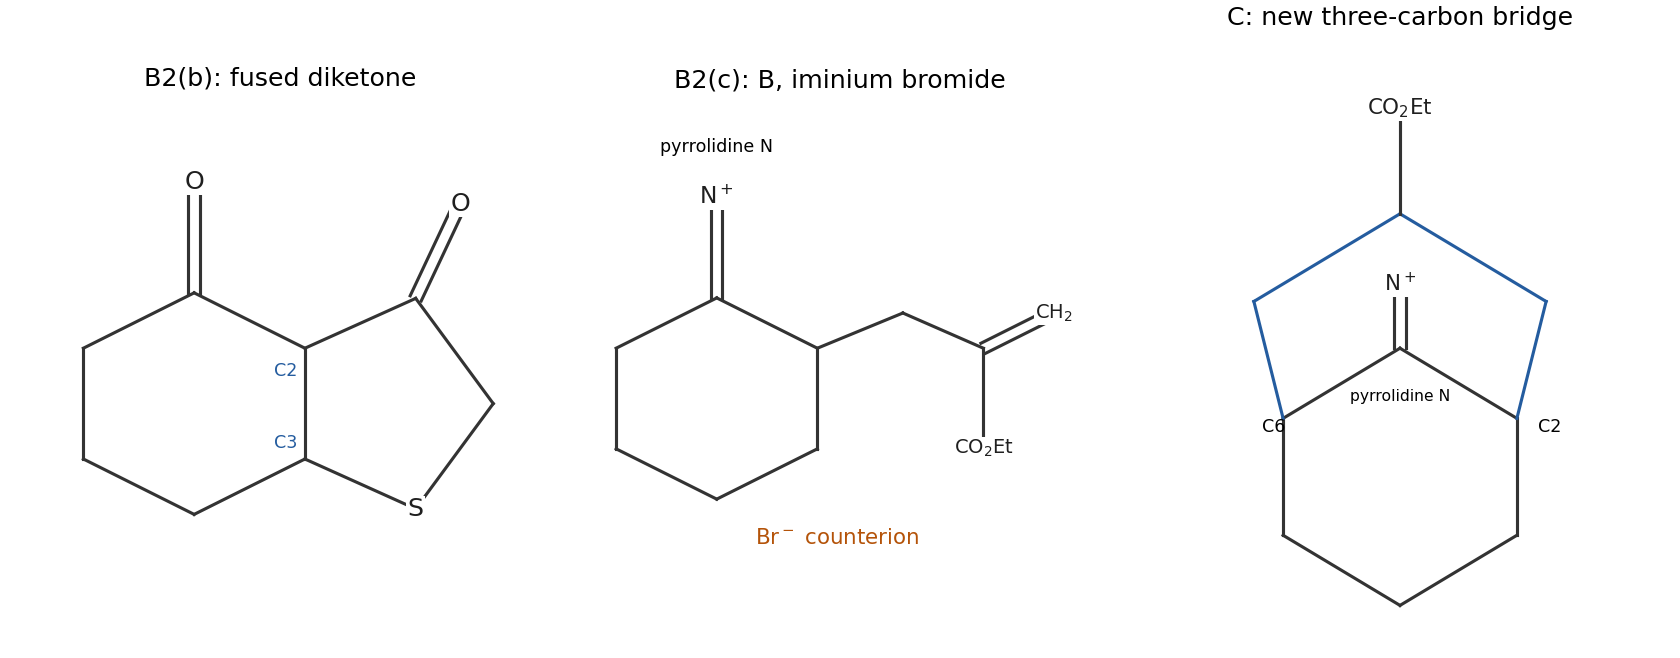
Label the enamine carbon bonded to nitrogen C1 and its alkene partner C2; C6 is the other carbon next to C1. B is the C-alkylated iminium ion bromide. Its new substituent at C2 is , while C1 is double-bonded to the pyrrolidine nitrogen, which bears positive charge. bromide is the counterion, accounting for Br in the empirical formula without requiring a retained C–Br bond.
The C2 end of the enamine attacks the bromomethyl carbon by displacement. Simultaneously, donation of the nitrogen lone pair creates the C1=N bond. This is ordinary enamine C-alkylation, not N-alkylation.
proton transfer removes an alpha proton at C6, converting the iminium ion into the enamine regioisomer C1=C6. That enamine attacks the terminal of the tethered acrylate in an intramolecular Michael addition. nitrogen donation restores the iminium ion, and protonation of the ester enolate completes the sequence.
The new bridge is . Together with the original ring and its C1 bridge this produces the bicyclononane iminium ion skeleton in C. It retains the pyrrolidine group and the ester; hydrolysis to a ketone is not part of the displayed reaction. The second panel of the original annulation figure shows the chemical atom connectivity and distinguishes ionic bromide in B from a covalent bromine substituent.
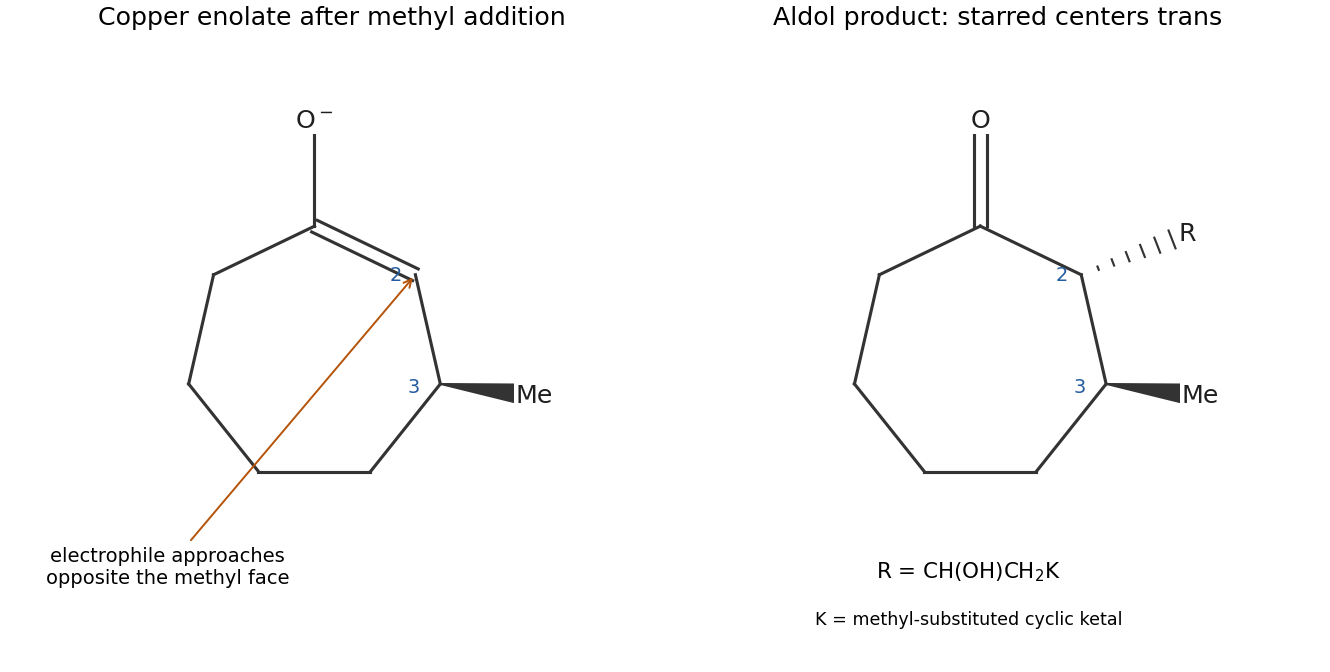
copper(I) converts the methyl grignard reagent into a softer organocopper nucleophile. Methyl undergoes 1,4 addition to the beta carbon of the cycloheptenone, giving a metal enolate rather than direct 1,2 addition to the ketone carbonyl.
The carbon nucleophilic site of this enolate, at the alpha carbon, attacks the aldehyde carbonyl in the second reagent. Its C=O electrons move to oxygen; protonation of the resulting alkoxide gives the beta-hydroxy ketone side chain. The cyclic acetal is a protected ketone and survives these nucleophilic conditions.
The beta methyl group shields one face of the cyclic enolate. Aldol bond formation from the less crowded opposite face places the new alpha side chain trans to the beta methyl group. Thus the two starred ring centers have an anti relationship; the two mirror-related products are possible from achiral starting materials and reagents. This argument does not assign the separate, unstarred alcohol center, for which the question gives no specified configuration.
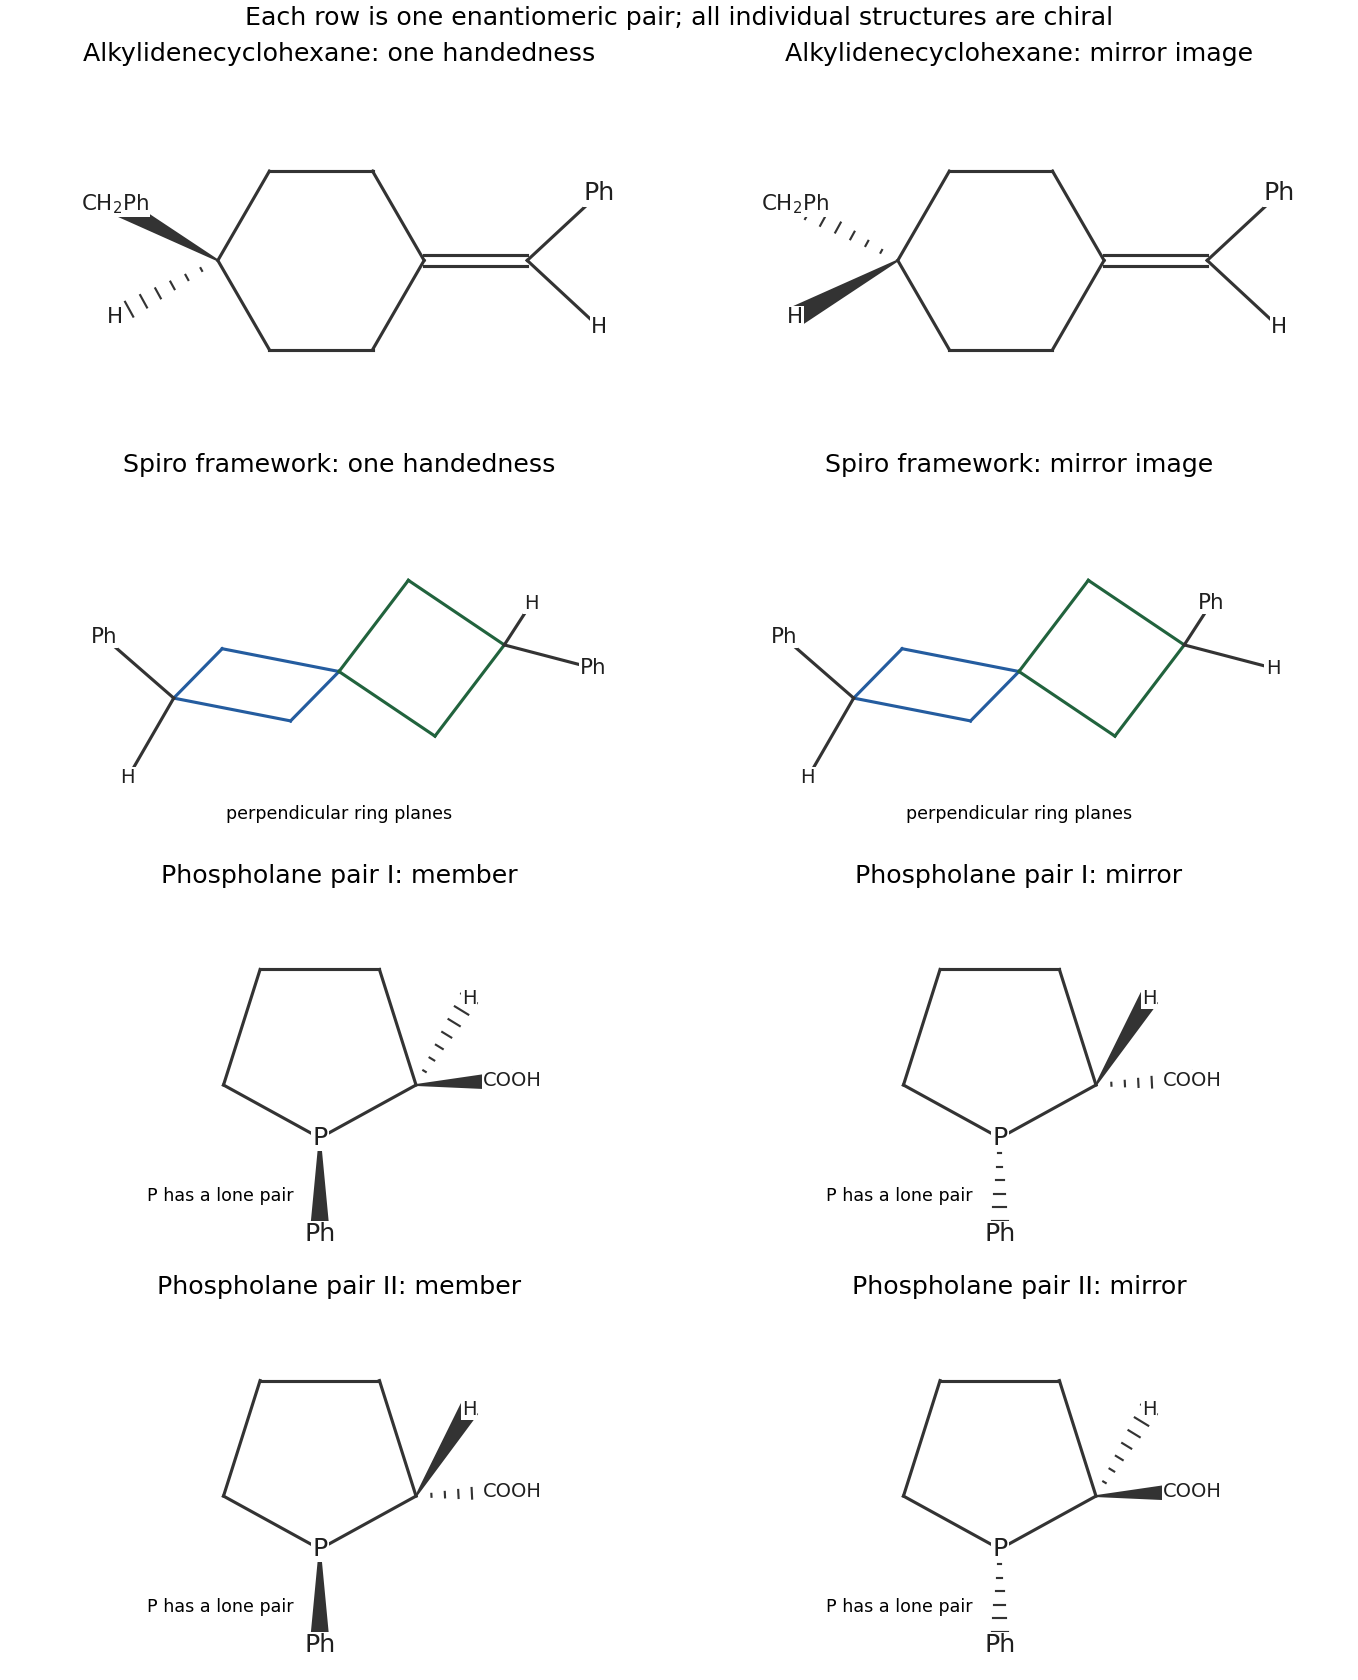
There are two, two and four stereoisomers, respectively; all eight individual stereoisomers are chiral. The drawings below give the mirror pairs. Solid and hashed wedges distinguish opposite faces; ordinary changes between conformers are not counted again.
- The benzyl-substituted alkylidenecyclohexane has an axial stereogenic element. At one end the exocyclic alkene presents Ph and H; at the other end the substituted ring carbon presents benzyl and H. The ring constrains the intervening paths, so exchanging one end pair gives the nonsuperposable mirror image. There is one enantiomeric pair, rather than an independent ordinary pair multiplied by a carbon-center count.
- The diphenylspiro compound also has one axial chirality. The two four-membered rings occupy different, approximately perpendicular planes. Their remote Ph/H substitutions distinguish the two handed arrangements. It gives one enantiomeric pair and no meso form. axial chirality of this spiro framework is experimentally established, for example by resolution of its 2,6-dicarboxylic-acid derivatives.[ref]Enantiopure spiro[3.3]heptane-2,6-dicarboxylic acid by Hong-Zhi Tang and others. Abstract, isolation of enantiopure spiro derivatives
- The phospholane has a stereogenic carbon at the COOH-bearing position and a pyramidal phosphorus stereocenter. The phosphorus has three different carbon ligands and a lone pair: its two ring paths differ because only one encounters the substituted carbon immediately. The four configurations are , , and . The first and last form a mirror pair, as do the middle two. carbon and phosphorus cannot be interchanged by a symmetry operation, so there is no meso reduction in the count.
Unlike the usual rapidly inverting trialkylamine, a pyramidal phosphine can retain a phosphorus configuration on an ordinary observational timescale; its two phosphorus configurations must therefore be included.
Both reactions are E2 eliminations requiring an axial chloride and an axial beta hydrogen in an antiperiplanar conformation relative to the C–Cl bond. The reacting C–H bond supplies the alkene bond as the C–Cl bond breaks; hydroxide accepts the proton.
For the trans-1,2 substrate, the low-energy chair has methyl and chlorine both equatorial. ring inversion is needed to place chlorine axial, but this also places methyl axial. The population of the reactive chair is consequently small, explaining the slow reaction. At the methyl-bearing neighboring carbon the axial position is occupied by methyl, not hydrogen. Only the other beta carbon has the needed anti axial H, so elimination gives 3-methylcyclohexene rather than the more substituted alkene.
For the cis substrate, a favorable chair has methyl equatorial and chlorine axial. It can react without paying the axial-methyl penalty, explaining the faster rate. Both adjacent carbons now supply an anti axial hydrogen. Removing the H on the methyl-bearing carbon gives 1-methylcyclohexene, the major product; removing the other gives 3-methylcyclohexene, the minor product. The former has the more substituted double bond.
Thus alkene stability selects between accessible pathways in the cis case, but it cannot create the missing antiperiplanar conformation H in the trans case.
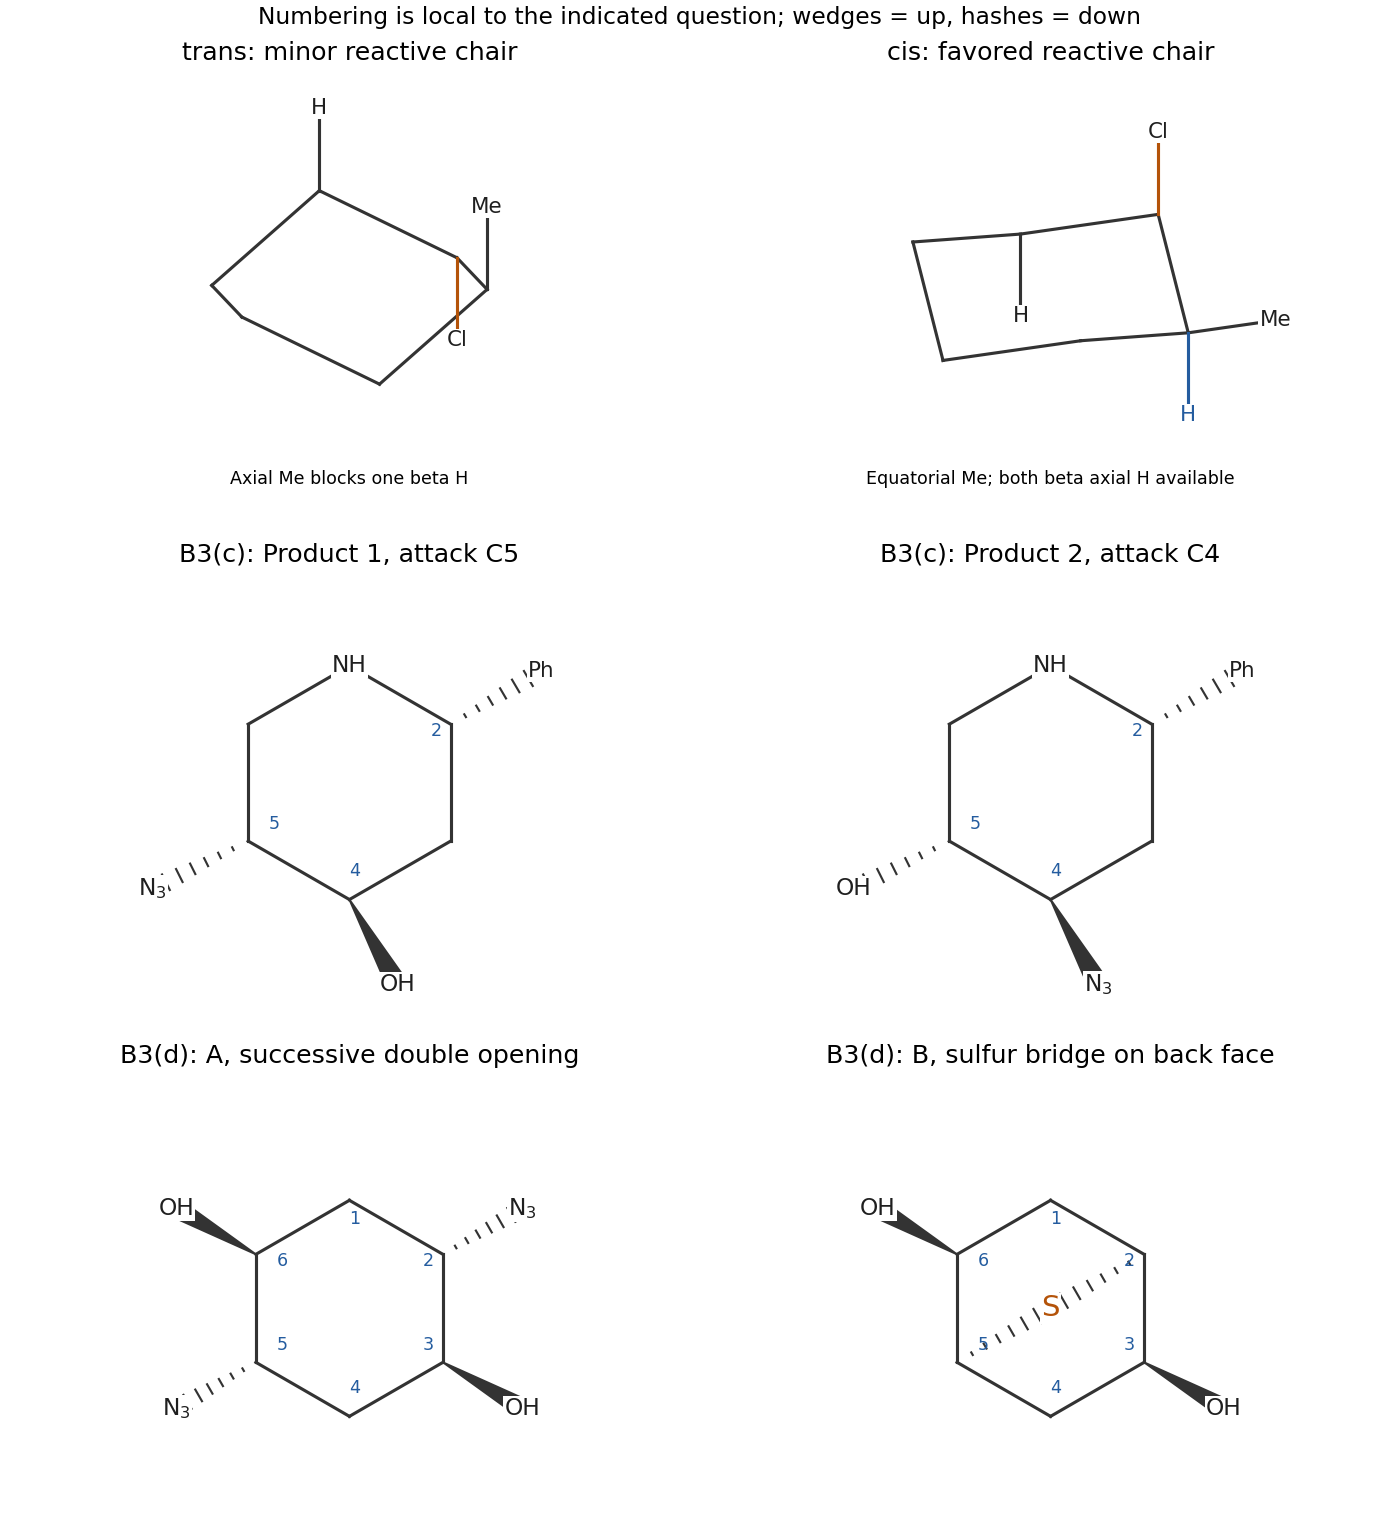


Number the nitrogen N1, the phenyl-bearing carbon C2, and then continue through C3–C6 around the piperidine. Call the solid-wedge epoxide face up; the phenyl group is down in both starting structures. The bulky phenyl group favors a pseudoequatorial orientation.
epoxide opening follows the Fürst-Plattner preference for a chair-like, trans-diaxial opening trajectory. Attack at the two epoxide carbons is not equivalent: the favored route evolves toward an ordinary chair, whereas the alternative demands a less favorable twist/boat-like distortion. Such conformational control of epoxy-2-arylpiperidine opening is supported by synthetic studies.[ref]
1,3-Allylic Strain as a Strategic Diversification Element for Constructing Libraries of Substituted 2-Arylpiperidines by Thomas C. Coombs and others. Scheme 1, chair-like trans-diaxial epoxide opening; applied here with the free-NH conformer
For the first epoxide, azide attacks C5 from down, breaking the C5–O bond. The residual C4–O bond is retained, so work-up gives C5–N3 down and C4–OH up, with the original C2–Ph down. For the second, down-face epoxide, azide attacks C4 from up, leaving C4–N3 up and C5–OH down. These assignments are shown in the middle row of the original stereochemical map.
In each mechanism an azide terminal nitrogen attacks the epoxide carbon, the C–O bonding pair goes to oxygen, and protonation gives the alcohol. Each attacked center inverts, and the azide and OH are trans. The phenyl-bearing center is unchanged. Reversing the epoxide face changes which carbon has the favored axial attack trajectory; a steric rule based only on distance from phenyl misses that conformational effect.
Number the cyclohexane consecutively, putting the top methylene group at C1 and the two original epoxides at C2/C3 and C5/C6. Both epoxide oxygens are up.
A is a diazidocyclohexanediol, with each epoxide replaced by one azide and one alcohol. A standard successive chair-favored opening gives N3 down at C2 and C5, OH up at C3 and C6. This is the 1,4-diazido/1,4-diol connectivity drawn in the bottom row of the original map.
For each opening, azide attacks from the face opposite oxygen, breaking one C–O bond with inversion at the attacked carbon. protonation of the two resulting alkoxides gives the two OH groups. After the first opening, the preferred conformation of the opened portion must be used for the second trans-diaxial opening; simply imposing one unchanged chair on both steps can select the wrong regioisomer.
The elemental change is addition of two groups and two work-up protons to , giving . The formula alone would not distinguish alternative positional isomers; the depicted assignment uses the ordinary successive-opening conformational model.
sulfide first opens one epoxide by backside attack, giving a carbon-bound thiolate and an alkoxide. Using the same numbering, take this first attack at C2: the C2–S bond is down and C3–O remains up.
The attached sulfur still bears a nucleophilic lone pair and negative charge. It attacks C5 of the second epoxide intramolecularly, breaking C5–O and closing a bridge between C2 and C5. Work-up protonates the oxygen anions at C3 and C6. Thus B is 7-thiabicycloheptane-2,5-diol, with both OH groups on the face opposite the sulfur bridge.
Both C–S bonds are formed by inversion at their attacked epoxide carbons. Because the original epoxides are cis, both new C–S bonds point to the same opposite face and one sulfur chemical atom can connect them. The 1,4 bridge gives two five-membered sulfur-containing circuits, unlike the more strained closure between nearby 1,3 positions.
The sulfur therefore replaces two separate external nucleophiles: two epoxides are opened but only one sulfur chemical atom is incorporated. Adding that sulfur and two work-up protons gives , consistent with B.
The alcohol oxygen attacks sulfur of methanesulfonyl chloride, chloride leaves, and triethylamine removes the O–H proton. This gives an O-mesylate with retention of the carbon configuration: the original carbon–oxygen bond has not been displaced.
An accessible chair places the mesylate axial. triethylamine then removes the anti axial hydrogen on the carbon between the ketone and mesylate. Its C–H bond forms the new C=C bond while the C–O bond expels mesylate. The result is the conjugated cyclohexenone.
The neighboring carbon on the acetonide side has its relevant axial position occupied by its oxygen substituent, rather than an anti axial hydrogen. The elimination toward the ketone is therefore favored both by the trans-diaxial geometry and by conjugation of the resulting alkene with C=O. The acetonide remains intact. No inversion of the original alcohol carbon is required during mesylation, and it becomes planar during elimination.
B is the major product: the newly formed OH is the dashed bond in the printed zigzag structure. Retain the existing solid-wedge OH and the configuration at the carbamate-bearing carbon.
zinc coordinates the ketone oxygen and the adjacent hydroxyl oxygen, forming a five-membered O–Zn–O chelate. This constrains rotation around the carbonyl–alpha-carbon bond. In the favored chelated conformation the bulky group shields one face of the ketone, while the alpha hydrogen leaves the other more accessible. hydride attacks from that less crowded face, the carbonyl bond opens onto oxygen, and aqueous work-up gives the second alcohol.
Mapping this Cram chelation model back onto the printed zigzag gives B, not A. Merely placing a new OH “on the same side” of an unrotated page drawing is not a valid chelation argument: the reactive chelate and that drawing use different conformations. Zinc-borohydride reductions of alpha-hydroxy ketones are experimentally known to favor the corresponding anti-1,2-diol family under chelation control.[ref]
The reagent combination can generate zinc borohydride and related zinc-associated reducing species. The stereochemical justification is coordination-controlled facial selectivity; it does not require identifying a unique solution aggregate from the reagent list. The quoted yield is not itself a measurement of a 95:5 diastereomeric ratio.
The first step is a vinylogous aldol condensation at C5 of the unsaturated lactam. Number nitrogen N1, the lactam carbonyl C2, and then the methoxy-substituted alkene C3=C4 and the methylene group C5.
hydroxide reversibly removes a C5 proton. The resulting vinylogous enolate is delocalized through C5–C4–C3–C2–O: it is not necessary to create an unstabilized isolated carbanion beside nitrogen. Its C5 nucleophilic contributor attacks acetone. protonation gives a C5-linked tertiary alcohol, . Further removal of the remaining C5 hydrogen and elimination through an aldol-dehydration sequence forms C5=C(Me)2, leaving the original methoxy enol ether in place.
In the second step, aqueous acid hydrolyses that enol ether to a ketone at C4. protonation at C3 gives an oxygen-stabilized cation at C4. water attacks C4, proton transfers form a hemiacetal-like intermediate, and loss of methanol followed by deprotonation produces C4=O and the C3 methylene group.
The lactam carbonyl and N–H group remain, as does the exocyclic isopropylidene group. This accounts for replacement of the C3=C4–OMe unit by C3H2–C4=O without inserting a new oxygen into the ring skeleton.
The hydroxyl group first binds aluminium, normally by deprotonation to an aluminium alkoxide with release of hydrogen. Coordination also brings aluminium close to the alkyne. directed trans-hydroalumination then puts hydride on the alkyne carbon proximal to CH2O and aluminium on the distal carbon.
One useful mechanistic description has external hydride attacking the coordinated alkyne opposite the aluminium-containing face. The resulting vinylaluminate is stabilized by a five-membered O–Al–C–C–CH2 chelate. This is not the ordinary undirected syn addition of a neutral dialkylaluminium hydride. Propargylic-alcohol-directed trans-hydroalumination with Red-Al gives vinylaluminate intermediates that can be intercepted by electrophiles.[ref]
trans-Hydroalumination and alkylation by Neil F. Langille and Timothy F. Jamison. Abstract, stereoselective vinylalanate formation from propargylic alcohols
iodine replaces the distal C–Al bond with C–I without scrambling the alkene geometry; aqueous work-up releases the alcohol. hydride and iodine are consequently trans across the alkene, while iodine and the CH2OH group are on the same side.
By Cahn-Ingold-Prelog priority rules priority the product is therefore : CH2OH outranks H at one alkene carbon and I outranks CH2R' at the other. Describing the two carbon chains as trans would not make this an alkene, because the iodine changes which substituent has higher priority.
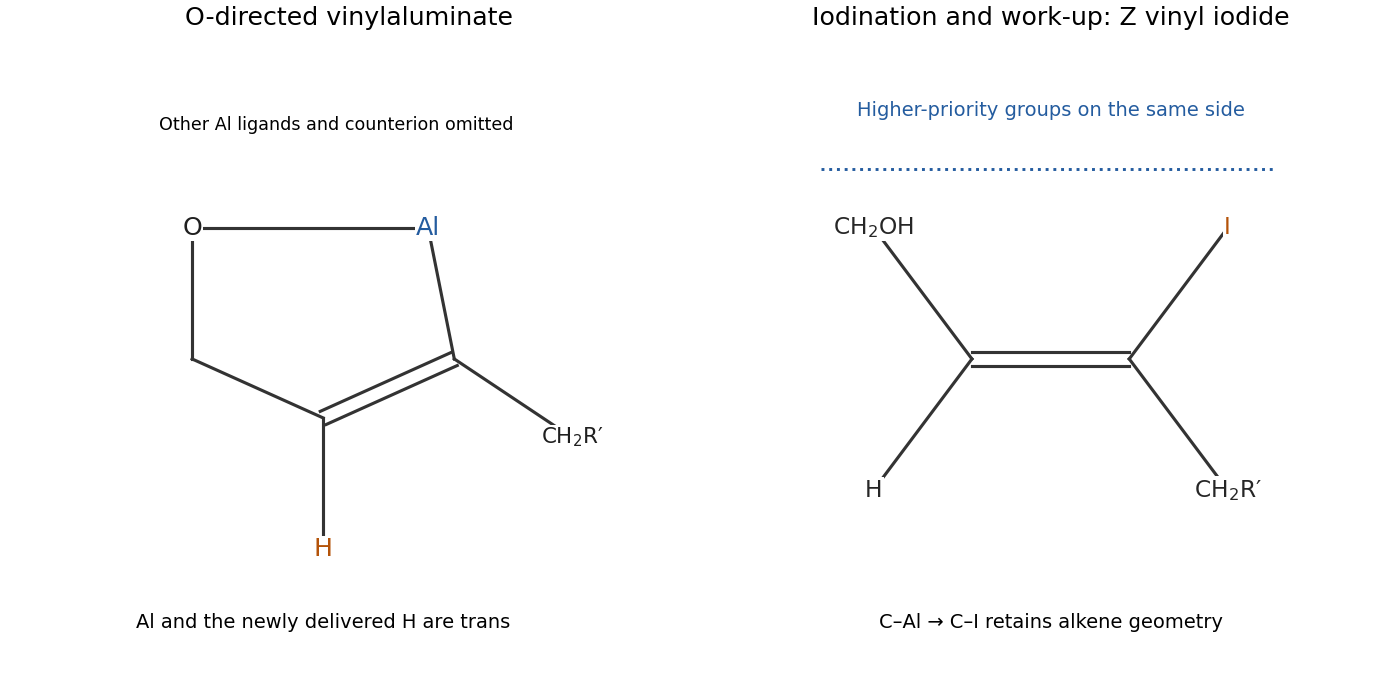
Directed anti hydroalumination, stereoretentive iodination and the Z priority assignment
. Details. 
TODO: user-deferred biology. This biological subpart is intentionally left unsolved under the current nonbiology-only scope.
Two molecules of the carboxylic acid form a cyclic carboxylic-acid dimer. Let Ar denote the meta-methylphenyl group. One molecule donates its OH hydrogen to the other's carbonyl oxygen, and the second donates back:
The two contacts close an eight-membered hydrogen-bonded ring, shown explicitly below.
The two contacts close an eight-membered hydrogen-bonded ring, shown explicitly below.
Both molecules therefore act once as hydrogen-bond donor and once as hydrogen-bond acceptor, giving two O–H···O=C contacts. This is the conventional self-complementary dimer motif; the schematic is not a prediction of every weaker contact in an unprovided crystal structure.
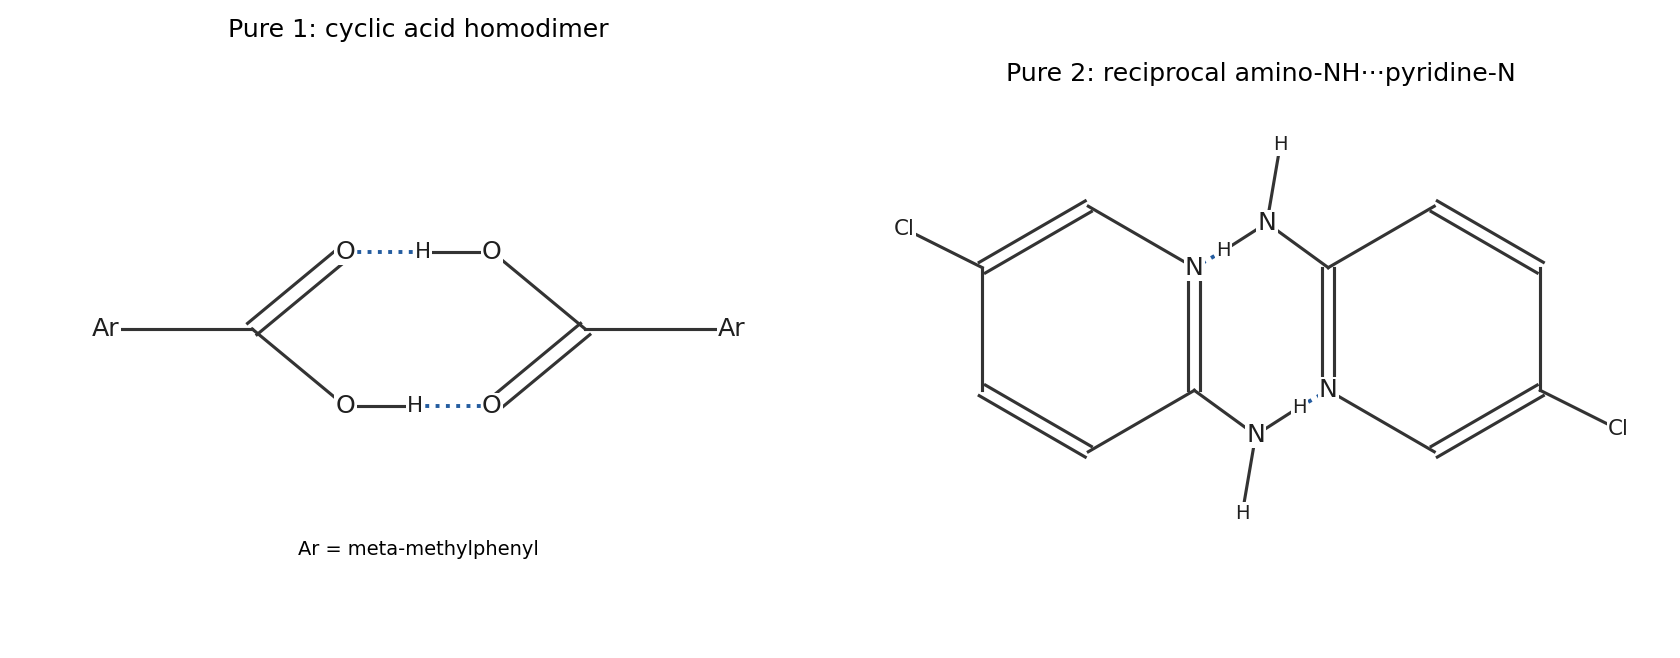
Original hydrogen-bonding structures for the two pure-compound homodimers
. Details. 
Two molecules of the aminopyridine form a self-complementary dimer through two N–H···N contacts. In each direction, one exocyclic amino hydrogen is the donor and the pyridine ring nitrogen of the other molecule is the acceptor.
Orient the two amino/pyridine-N edges toward one another, as in the second panel of the original homodimer drawing. Each molecule donates one NH and accepts at one ring N; the second amino H remains available for additional packing contacts. The chlorine substituent is retained but is not one of the two principal donor–acceptor sites.
The strongest donor belongs to the acid, , and the strongest acceptor to the aminopyridine, . The cocrystal pairs the strongest donor with the strongest acceptor, while the reciprocal amino-NH···carbonyl contact closes a complementary two-contact motif.
Using the simple favorable-contact score , one heterodimer givesThe self-associated comparison, normalized to the same two molecules and two hydrogen bonds, is half an acid dimer plus half an aminopyridine dimer:ThusIn a model where bond free energy contains , this is a favorable energy change. No bonds have been artificially added to one side of the comparison: both alternatives contain two contacts per acid/aminopyridine pair.
The calculation rationalizes the hydrogen-bond preference. It is not a complete crystal free energy: packing, additional contacts, entropy and possible solvent effects also matter. Without a specified calibration, the numerical score should not be promoted to a measured cocrystallization free energy.
TODO: user-deferred biology. This biological subpart is intentionally left unsolved under the current nonbiology-only scope.
Articles by others on the same topic
There are currently no matching articles.