For a monocyclic, approximately planar, continuously conjugated pi system, Hückel's rule predicts aromatic closed-shell filling for pi electrons, with . Every atom on the circuit must have an appropriately oriented orbital; counting double bonds without checking pi conjugation is insufficient.
The two outer benzene rings are clearly aromatic; the central pyrone ring has a weaker, polarized six-pi aromatic contribution. Each outer ring has the familiar six-electron circuit. For the central ring, a useful charge-separated contributor moves the exocyclic carbonyl pi pair onto oxygen and donates one endocyclic oxygen lone pair into the ring. Its four fused carbon p centres contribute four electrons and the ring-oxygen lone pair supplies two: a six-pi, pyrylium-like circuit, with positive charge on the ring oxygen and negative charge on the carbonyl oxygen. This does not make three independent benzene rings of equal aromatic stabilization; localized carbonyl bonding and the fused benzene circuits remain important contributors.
The three oxygens play different roles. The endocyclic ether oxygen donates a p-type lone pair to pi conjugation. The carbonyl oxygen accepts pi density in the charge-separated contributor. The phenolic oxygen donates into the outer ring as a substituent. Its in-plane lone pair is not an extra pair to add to every ring's electron count. In particular, neither the external C=O pair nor both lone pairs on a ring oxygen should be counted as independent extra ring electrons. Structural studies of xanthone support a largely planar conjugated framework, while the division into strongly aromatic outer rings and a more polarized central ring is an electronic-structure interpretation.[ref]
Structure of xanthone by Shigeki Onuma and others. 1990, pp. 1725–1727, molecular geometry
B is 1,3-dibromo-2-hydroxyxanthone: both bromines are on the hydroxy-substituted outer ring, at its two available positions ortho to OH. Define C2 as the OH-bearing carbon; C1 lies between it and the carbonyl-side ring junction, and C3 lies on its other side. These labels specify the substitution independently of the orientation of the printed drawing.
Phenolic lone pair donation strongly activates its ortho and para positions towards electrophilic substitution. The para position is a fused junction and has no replaceable hydrogen, leaving C1 and C3 as the two directly activated sites. C1 also benefits from the conjugative influence of the ring ether oxygen. Although other positions receive some activation from that oxygen, they lack the stronger local phenolic activation of the selected pair. Excess bromine can therefore replace both ortho hydrogens. Replacing two hydrogens in gives , consistent with the stated composition. Addition across an outer-ring double bond would sacrifice aromaticity and would not account for the hydrogen loss.
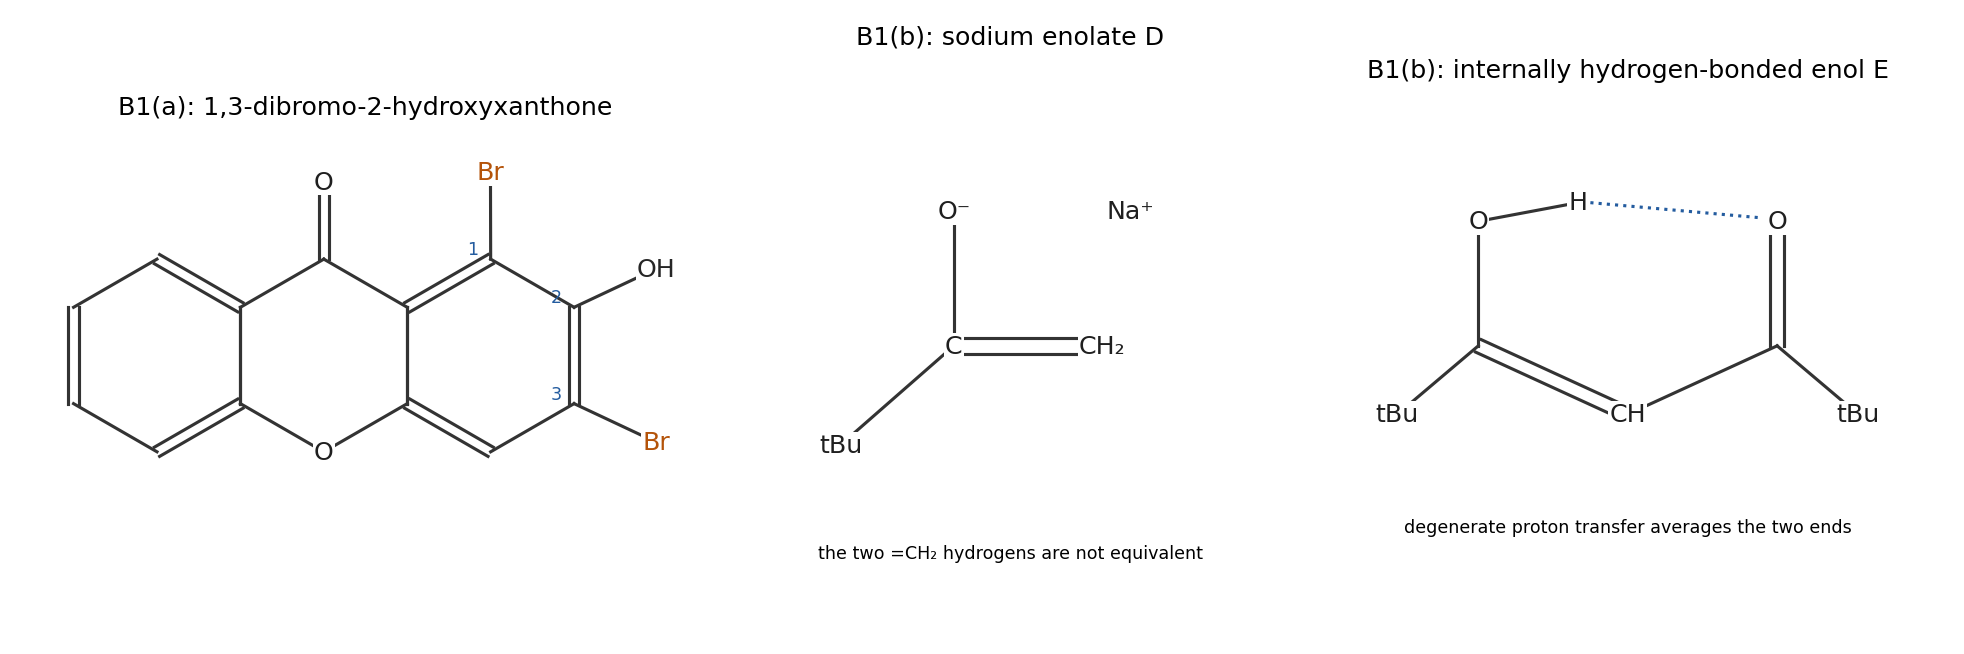
Original solution structures of the dibrominated xanthone, sodium enolate D and hydrogen-bonded enol E
. Details. 
The activated aromatic pi system polarizes bromine and attacks its electrophilic end while Br–Br cleaves. This makes a C–Br bond at C1 or C3 and gives a Wheland intermediate. Donation from phenolic oxygen stabilizes contributors with positive charge at its attached ring carbon. acetate or bromide then removes the proton from the attacked carbon; the C–H bond electrons restore the aromatic pi system. Repetition at the other activated ortho site gives the dibrominated product.
Each step replaces C–H by C–Br and restores aromaticity. No radical initiator or benzylic abstraction is needed, and no Lewis acid catalyst should be invented for this already activated phenolic substrate. The overall substitution balance is .
Write . D is the sodium enolate . Its two resonances in the alkene-proton region belong to the two hydrogens of the terminal methylene group. One is cis to oxygen and the other trans to oxygen, so they are not related by a symmetry operation even though both belong to the same carbon. Replacing one or the other gives different alkene geometries, so they are diastereotopic. Their weak or unresolved geminal coupling is consistent with the reported singlet descriptions. Without an additional spatial assignment, it is not justified to decide uniquely which numerical shift belongs to which of those two orientations. The nine-proton singlet is the tert-butyl group. The formula follows from removing one alpha hydrogen from pinacolone.
E is the enol-dominated 1,3-diketone , observed as . The very downfield broad signal belongs to the enol OH in a six-membered intramolecular O–H···O chelate. The one-proton signal in the vinyl region is its central enol CH; a predominantly diketo structure would instead require a two-proton central CH2 signal and no enol OH. The eighteen-proton singlet accounts for both tert-butyl groups. Rapid exchange between the two degenerate enol orientations can average their different local environments; accidental unresolved equality cannot be excluded by the supplied spectrum alone. Thus one should not infer a permanently symmetric single enol Lewis structure from that singlet. The total integration is , agreeing with .
amide ion removes a methyl-side alpha proton of pinacolone. The C–H electrons form the enolate C=C bond as the carbonyl pi pair moves onto oxygen; ammonia is formed. The tert-butyl-side alpha carbon has no hydrogen, fixing the ordinary enolate regiochemistry.
The carbon end of D attacks the acyl chloride carbonyl carbon. Move the C=O pi pair onto oxygen, then collapse the tetrahedral intermediate with chloride departure. This nucleophilic acyl substitution gives and constructs the bond between the original enolate methylene and the incoming carbonyl. The observed product requires carbon acylation, rather than an O-acyl enol ester. A further equivalent of D removes the much more acidic central proton of the new 1,3-diketone, generating its extensively delocalized enolate and returning that second D equivalent to pinacolone. Aqueous acid workup protonates the product enolate; keto–enol equilibration gives the conjugated, internally stabilized by a hydrogen bond enol observed by NMR. The acyl chloride contributes its entire pivaloyl group; neither tert-butyl group is lost.
One equivalent supplies carbon; the other acts as a base for the product. The newly formed 1,3-diketone is appreciably more acidic than the ketone from which D was prepared. proton transfer from its central CH2 to a second enolate equivalent is therefore favourable. Product-enolate stabilization also helps drive the acyl-substitution sequence towards completion.
The second enolate is consequently consumed as base, yielding pinacolone, rather than supplying a second carbon fragment to E. acid workup reverses the product deprotonation. Ignoring this acid–base step would predict that a single equivalent of D is sufficient to consume every equivalent of acyl chloride, despite the product immediately quenching part of the available reagent.
The five-carbon unit is initially an open chain, not a completed carbocycle. Label it , with phenyl-bearing , carbonyl , amide-substituted , and the two ethylthio groups on . Its double bonds are initially and . Let be the nitro-bearing carbon of nitroethane; its methyl remains a substituent throughout.
DBU removes an alpha proton from nitroethane to give a nitronate. carbon attack at the Michael acceptor forms , with electron redistribution through onto the carbonyl oxygen. proton transfer restores the ketone. Removing the remaining X–H proton gives another carbon-nucleophilic nitronate. It attacks the other conjugated end, , intramolecularly, closing the six-membered circuit . Addition to the ketene dithioacetal and ethylthiolate elimination restore the bond, leaving one SEt group at .
base-assisted loss of the proton at , with C–NO2 cleavage at X, creates and expels nitrite. Enolization at supplies the third aromatic double bond and the phenolic OH. proton transfers to the leaving anions complete the acid–base balance. nitroethane supplies the new ring carbon X and its methyl substituent; its nitro group is eliminated. Aromatization, rather than a separate oxidation of an intact six-membered starting ring, provides the final driving force. This is a proposed stepwise nitroalkane benzannulation; related DBU-promoted tandem Michael addition/cyclization/aromatization has direct primary precedent.[ref]
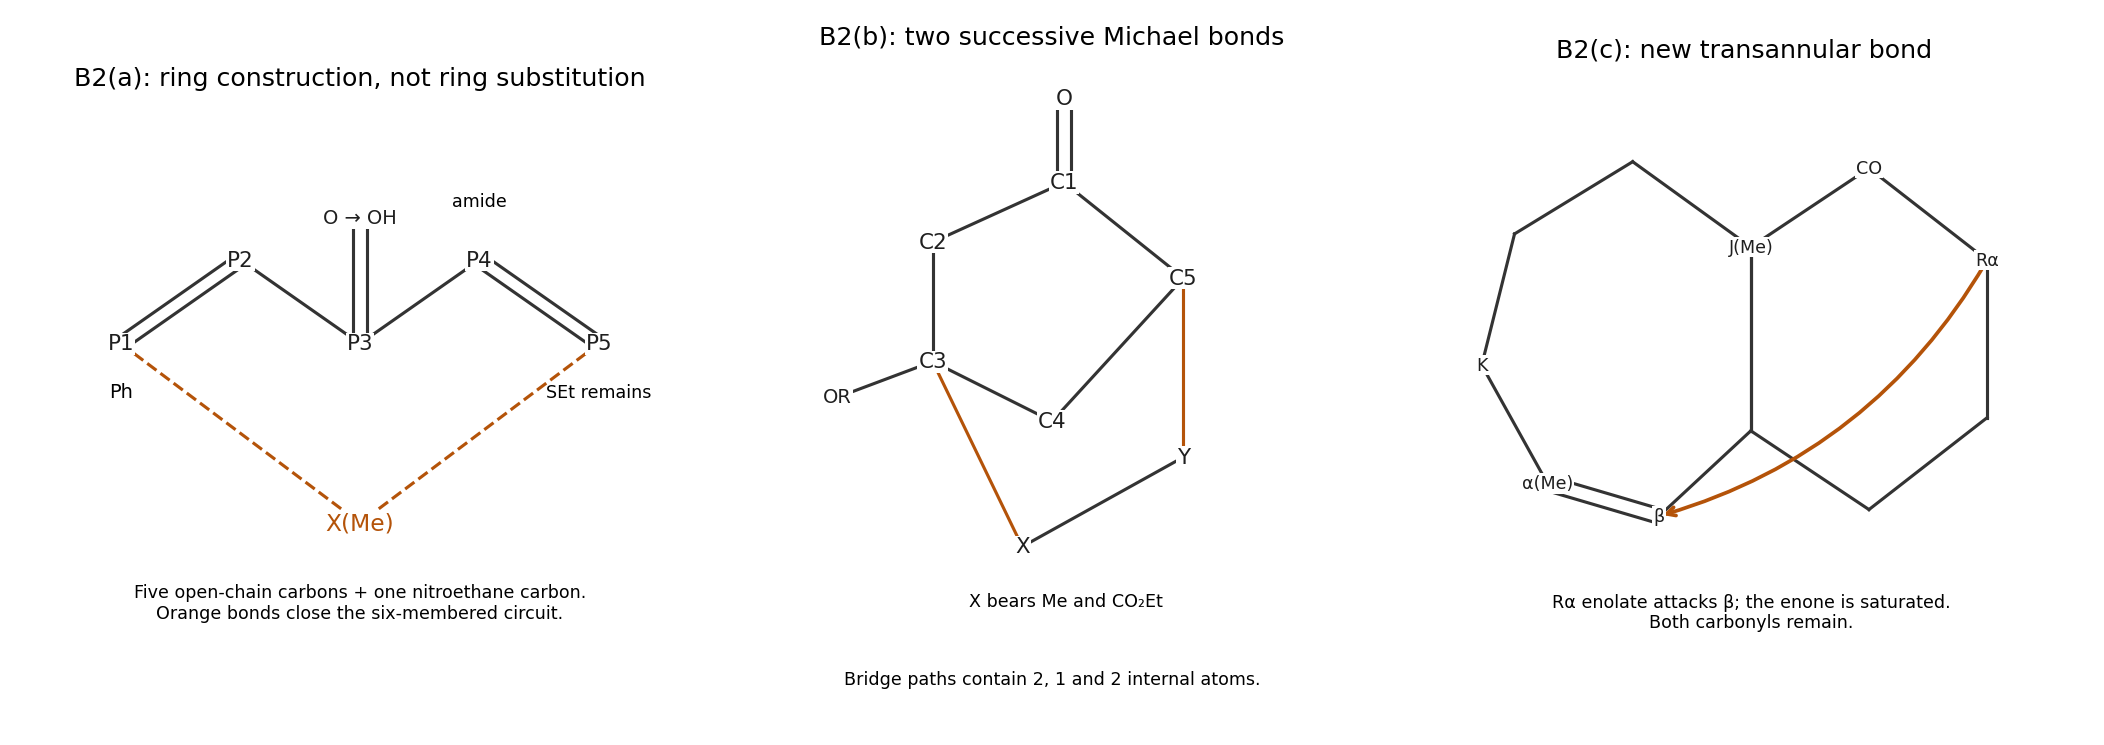
Original atom maps for the open-chain benzannulation, the two bonds of the norbornanone-forming tandem Michael addition, and the intramolecular Michael closure
. Details. 
Number the cyclopentenone carbonyl carbon , its alkene carbons and with OR at , and its saturated carbons and . The ester alkene has alpha carbon , bearing Me and CO2Et, and beta carbon , bearing the protected carbon substituent. lithium diisopropylamide generates the ordinary ketone enolate at the saturated alpha carbon . The acetonide is retained.
The carbon end of this enolate attacks ester beta carbon Y. Its pi system shifts through X onto the ester oxygen, forming the first new bond and an ester enolate at X. That enolate now attacks the original cyclopentenone beta carbon intramolecularly, giving the second new bond . The original enone electrons move onto its ketone oxygen, and protonation of the final ketone enolate completes the product. The OR substituent is retained at the new bridgehead; it need not leave in either Michael addition.
The atom map provides a useful structural check. Between bridgeheads and , the three paths are , , and . Their numbers of internal atoms are 2, 1 and 2, establishing the norbornane skeleton. Both original carbonyls survive and both initial alkenes are consumed. The two successive conjugate additions give this atom map without requiring a concerted cycloaddition.
The new bridge is made by an intramolecular Michael addition. On the cyclohexanone side, the methyl-bearing fused alpha carbon is quaternary and cannot be deprotonated there. triethylamine can instead generate a small equilibrium population of the enolate at the other, methylene alpha carbon. Its carbon attacks the beta carbon of the enone in the seven-membered ring. The enone pi electrons move towards its carbonyl and then onto oxygen, creating the new bridge and a second enolate.
protonation of that enolate at the former methyl-substituted enone alpha carbon restores the left-hand ketone. The initial enone C=C has therefore become a saturated C–C bond while one new transannular C–C bond forms. Both ketones are retained, and the initial two-ring framework becomes a three-ring framework without changing the total index of hydrogen deficiency. The methyl originally at the fused quaternary centre does not migrate; the other methyl ends up on a protonated, formerly unsaturated carbon. A strong-base concentration large enough for quantitative enolate generation is unnecessary for a favourable intramolecular trapping reaction, and neither an aldol dehydration nor a methyl shift is needed.
Let and number the carbon substituent from oxygen: . sodium hydride removes a proton from C3, alpha to the nitrile, forming hydrogen and a nitrile-stabilized carbon anion. That carbon attacks the nitroaryl ipso carbon bearing O. The tether closes a five-membered cyclic Meisenheimer intermediate with sequence C3–C2–C1–O–aryl ipso carbon. The para nitro group stabilizes the negative charge generated by temporary loss of aromaticity.
Rearomatization breaks the aryl–O bond, releasing the tether oxygen as an alkoxide while retaining the new aryl–C3 bond. protonation gives . This is a Truce–Smiles rearrangement: the aryl group migrates from oxygen to carbon. The nitrile remains attached to its original side-chain carbon; the original ether oxygen becomes the terminal alcohol oxygen. The substrate and product have identical atomic composition. Ordinary substitution at a saturated chain carbon would not produce this aryl–carbon bond or restore the terminal OH.
Number the OH-bearing carbon C1 and the adjacent brominated carbon C2. In the trans bromohydrin, base generates an alkoxide. A chair conformation can place O at C1 and Br at C2 trans-diaxial, aligning the oxygen nucleophile for backside attack at C2. Intramolecular SN2 closure expels bromide and inverts C2: the retained C1–O bond and the newly formed C2–O bond are both on the original OH face, giving the epoxide shown. ring inversion can provide the reactive chair even if it is not the dominant ground-state conformer.
The cis bromohydrin cannot place OH and Br trans-diaxial: one is axial when the other is equatorial. In the chair with Br axial, the hydrogen on C1 can instead be antiperiplanar to C2–Br. An alkoxide-assisted semipinacol pathway moves the C1–H bonding pair to C2 while the oxygen lone pair forms C1=O and bromide departs. hydride migration gives the ketone without changing the carbon skeleton. The hydrogen is transferred to C2 rather than expelled as a free hydride. The original phenyl-bearing stereocentre is not involved in either bond-reorganization sequence. Configuration controls which properly aligned displacement or migration can occur; a flat ring drawing alone conceals that distinction.
protonation of the secondary OH permits water departure. The neighbouring phenylthio sulfur can assist this departure by forming a bond to the adjacent alcohol carbon, generating a thiiranium ion spanning that carbon and the original sulfur-bearing cyclohexane carbon. Label the latter Q and the former L. The temporary Q–S–L bridge transfers the phenylthio attachment from Q to L when its Q–S bond subsequently breaks. sulfur attack opposite the departing OH accounts for the new C–S stereochemistry without requiring a freely rotating carbocation.
For the protected terminal alcohol, no free tethered OH is available to trap the bridge. Removal of a beta hydrogen from a cyclohexane carbon adjacent to Q forms the ring alkene while Q–S breaks. sulfur remains bonded to L, giving the allylic phenyl thioether. acid is regenerated by the deprotonation, so the catalyst need not be consumed.
For the free terminal alcohol, its oxygen attacks Q intramolecularly, opening Q–S instead. Count the new ether ring: O–CH2–CH(Me)–L–Q, a five-membered ring sharing Q with the cyclohexane. deprotonation gives the spirocyclic ether. This is capture of the same neighbouring-group-stabilized intermediate rather than migration of the methyl group. The pre-existing methyl-bearing carbon retains its configuration; its wedge/dash appearance changes when the product is redrawn in a different ring projection. The two substrates differ in whether a suitably placed oxygen nucleophile is available for intramolecular trapping.
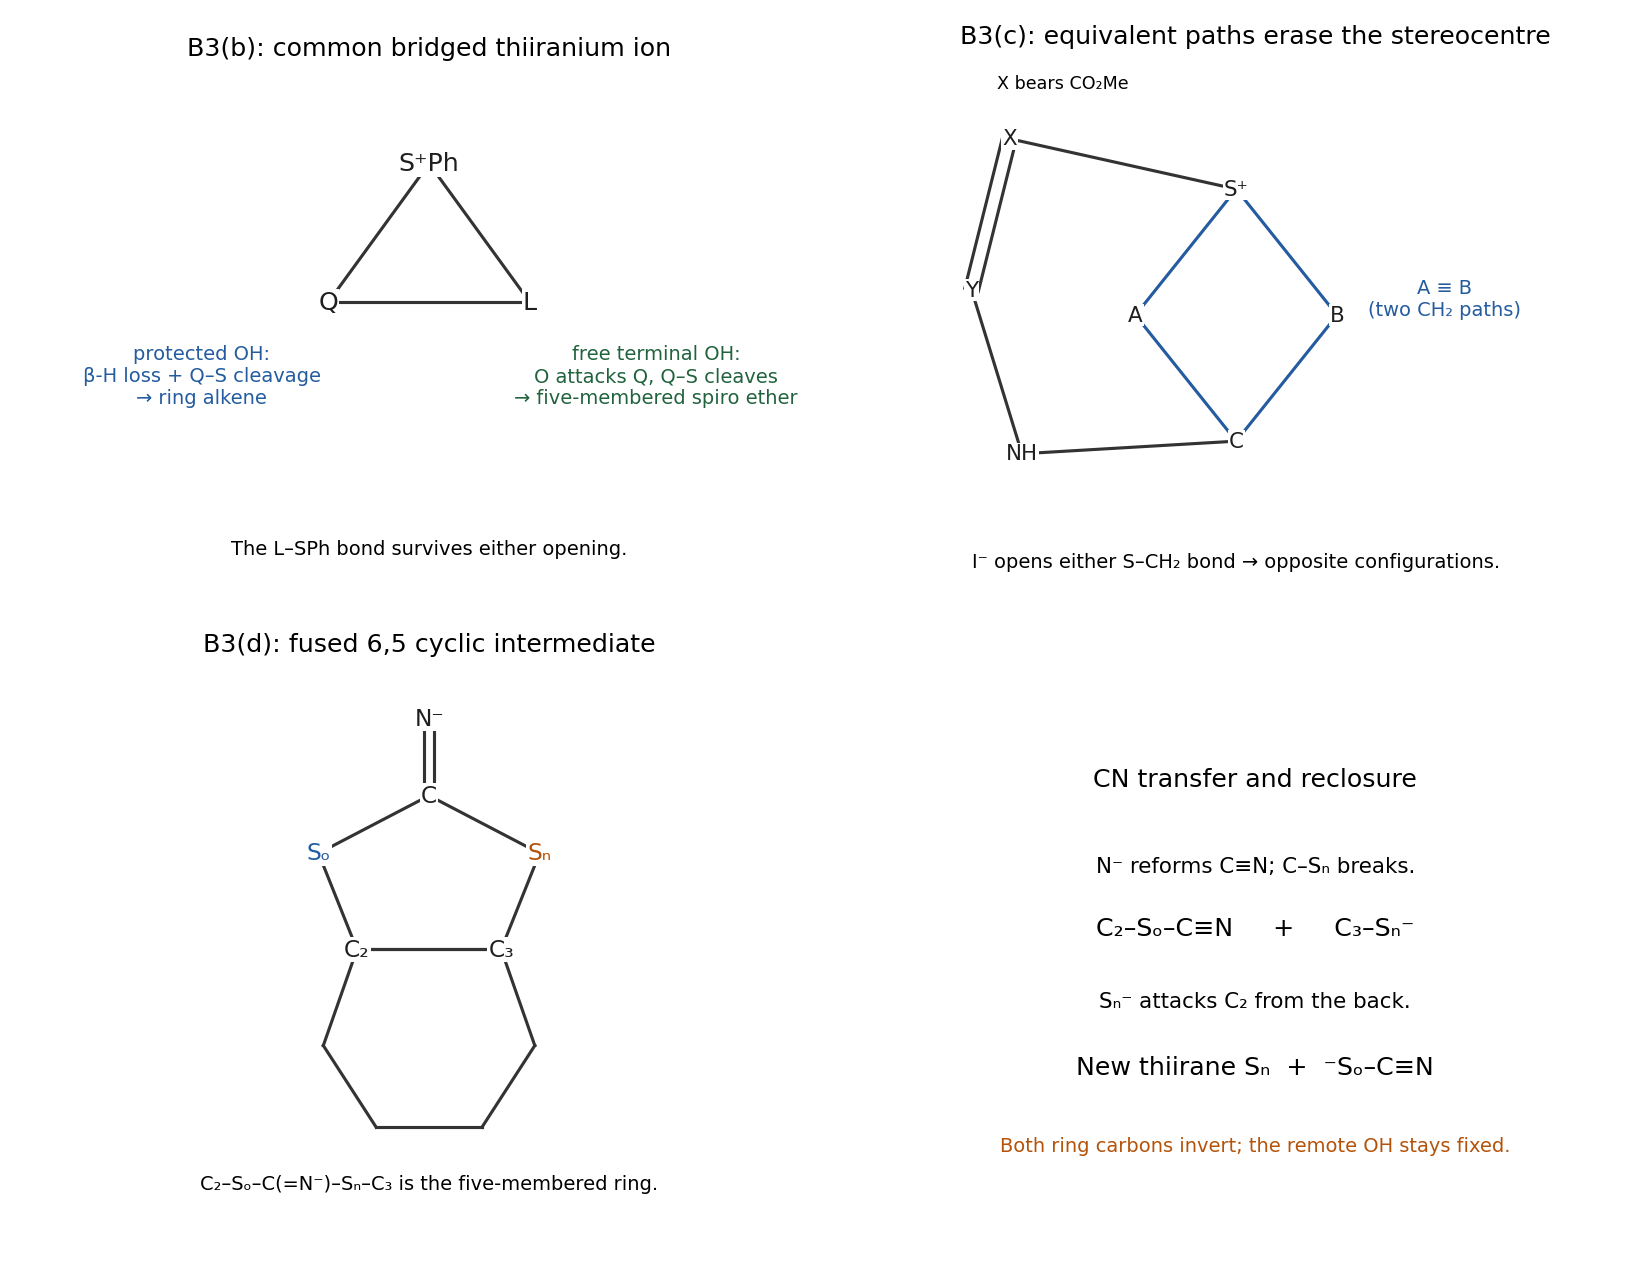
Original local atom maps of sulfur participation, the equivalent paths of the bridged thietanium ion, and the five-membered intermediate in thiirane sulfur exchange
. Details. 
Direct iodide attack at the terminal CH2–OMs would replace the leaving group without disturbing the adjacent stereocentre, so it cannot by itself explain racemization. sulfur participation produces an intermediate with two equivalent methylene paths. Label the original S-adjacent methylene A, the stereogenic carbon C, and its CH2–OMs substituent B. The thioether sulfur attacks B intramolecularly and expels mesylate, forming a four-membered thietanium circuit S–A–C–B–S. The rest of the original six-membered ring remains as the other path between S and C.
In this bridged sulfonium ion, A and B are both CH2 groups joining the same S and C centres. Exchanging them no longer changes the labelled molecular connectivity: the distinction that made C stereogenic has been removed. iodide can open the strained intermediate by backside attack at either A or B. Opening one S–CH2 bond gives one configuration of the six-membered product, while opening its equivalent partner exchanges which methylene is in the ring and which is the iodomethyl substituent, giving the opposite configuration.
The carbon iodinated in each opening is a methylene; racemization appears at the adjacent carbon through exchange of equivalent paths. It is not inversion at a CH2 stereocentre, nor does the mesylate carbon have to form a primary free carbocation. An achiral intermediate in an achiral medium gives equal mirror-related outcomes. nitrogen participation alone in a simple terminal displacement would not provide this specific equivalent-path explanation.
Distinguish the original thiirane sulfur from the sulfur supplied by thiocyanate. sulfur attack by at one thiirane carbon opens its C– bond with inversion. This produces a thiolate at the neighbouring carbon and a C––CN group at the attacked carbon. The OH-bearing remote carbon is unaffected.
The original thiolate now attacks the nitrile carbon intramolecularly. Shift one C≡N pi pair onto nitrogen. The resulting cyclic imidodithiocarbonate has the five-membered sequence ring carbon––C(=N)––adjacent ring carbon; its two carbon atoms are also part of the original six-membered carbocycle, exactly giving the hinted bicyclic intermediate.
Collapse can reform C≡N while breaking the nitrile-carbon– bond. This transfers the CN group to and generates as the thiolate on the other ring carbon. That thiolate attacks the remaining C– bond from the back, expelling and reclosing the thiirane. Each thiirane carbon has undergone one inversion, so the new sulfur bridge lies on the opposite face; the OH configuration is retained. The thiocyanate sulfur has entered the ring and the original ring sulfur has left as thiocyanate. This atom-labelled exchange mechanism explains how the same elemental formula can conceal sulfur replacement; isotope labelling would be needed to test that exchange experimentally. A single epimerization at the OH carbon would give the wrong stereochemical outcome.
A is the predicted major diastereomer. At the carbon adjacent to the ketone, take phenyl as the large group, vinyl as medium, and hydrogen as small. There is no adjacent heteroatom donor that would force a five-membered chelate. The appropriate first model is therefore the non-chelating Felkin-Anh model.
View along the carbonyl-carbon–stereocentre bond. Rotate to place the large alpha phenyl group approximately perpendicular to the carbonyl plane. Of the two resulting approaches opposite that large group, the favourable Bürgi-Dunitz trajectory passes nearer hydrogen rather than vinyl. Propyl transfer to the carbonyl and protonation of its oxygen give the configuration labelled A: in the printed projection the new OH projects forward and the propyl bond backward, while the original vinyl bond remains backward. Rotation used to construct a Newman projection must not be mistaken for inversion of the original stereocentre.
This is a relative-selectivity prediction, not a numerical product ratio. Simple attack on the face that happens to look less crowded in the initial flat drawing is unreliable because the relevant C–C single bond can rotate. B is the competing product from the less favourable approach beside vinyl. The model and its preference for the trajectory beside the small group are described in the institutional treatment.[ref]
Felkin-Anh and Cram Chelate by University of Windsor. pp. 1–2, section 1, nonpolar alpha substituents and the small-group trajectory
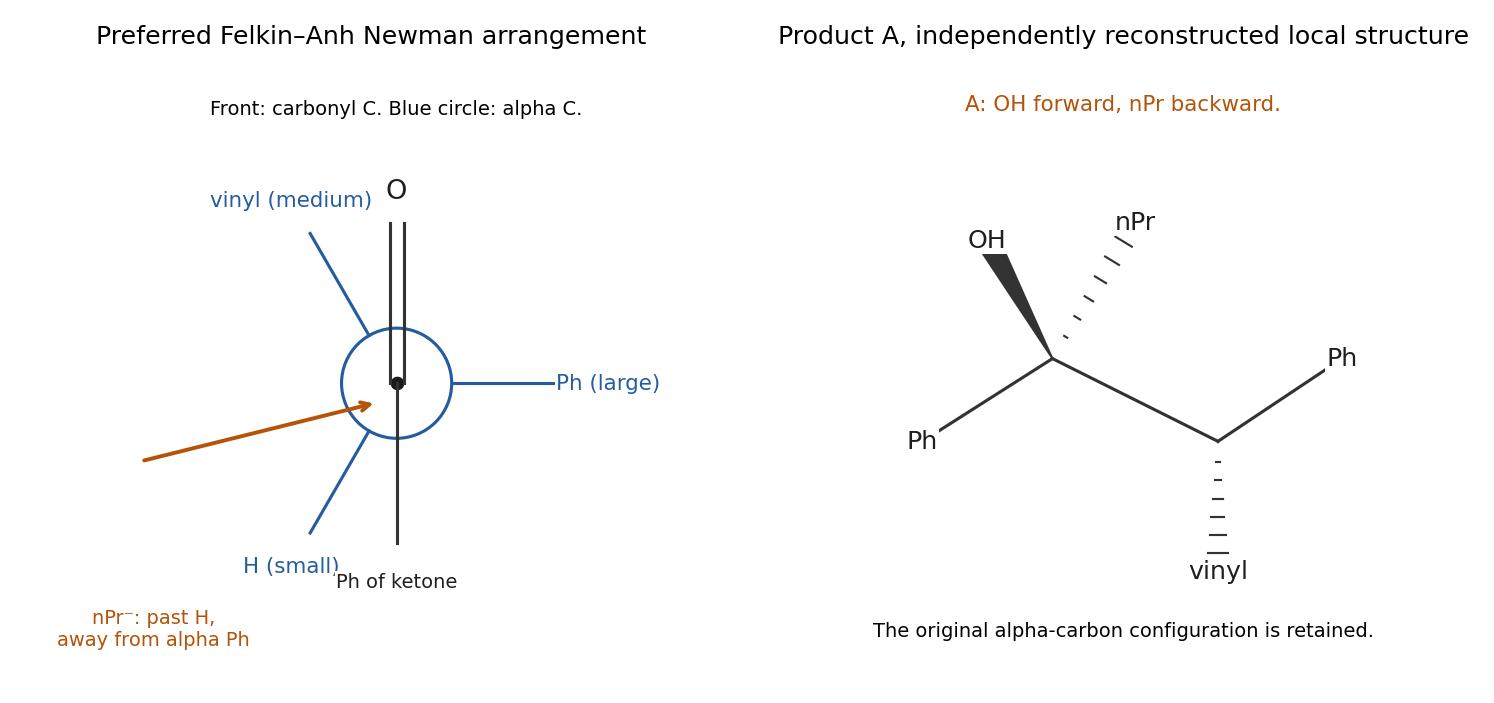
Original Newman construction of the preferred Felkin–Anh approach and the local stereochemistry corresponding to product A
. Details. 
The alkene attacks electrophilic mercury(II), with acetate departure as appropriate, to give a bridged mercurinium ion between the ring carbon and the exocyclic methylene. That bridge is polarized: the more substituted ring carbon can better stabilize developing positive charge than the terminal CH2 carbon. water attacks the ring carbon, opening the mercury bridge there, and deprotonation gives the tertiary alcohol with an exocyclic CH2–HgOAc substituent.
Subsequent sodium-borohydride demercuration replaces the terminal C–Hg bond by C–H. Thus the original exocyclic CH2 becomes Me, and the new OH remains on the substituted ring carbon. The result is Markovnikov hydration of the exocyclic alkene. There is no need for a freely equilibrating tertiary carbocation, and no rearrangement of the carbohydrate carbon skeleton or cleavage of the methyl acetal is required. The question does not require the mercury-stage facial selection or a detailed demercuration mechanism.
protonation activates the oxygen of the anomeric acetal bridge. Cleavage of its bond to the anomeric carbon releases that tether oxygen as the terminal primary alcohol, while the other ring oxygen stabilizes the developing anomeric electrophile by lone pair donation. The resulting oxocarbenium description has a nearly planar reactive anomeric centre; it need not be drawn as a long-lived free carbocation.
methanol attacks that centre, giving a protonated methyl acetal, and deprotonation regenerates acid. The former bridge becomes CH2OH; the anomeric substituent becomes OMe. The other C–O bonds and existing carbon configurations are retained. The ring can adopt a different chair once the bridge constraint is removed, so an apparent movement of substituents in the projection does not imply inversion at all the other carbons.
The depicted anomeric OMe is axial in the product chair. This geometry allows an endocyclic oxygen lone pair to align antiperiplanar to the exocyclic C–OMe bond, providing stabilization associated with the anomeric effect. Acidic conditions also permit cleavage and recapture at the anomeric centre, so selection is not simply backside displacement at an unchanged tetrahedral starting carbon. The anomeric effect rationalizes the specified axial product; the scheme alone gives neither its numerical equilibrium population nor evidence that every other anomer is absent.
triethylamine removes an alpha proton from nitromethane to generate a nitronate population. Its carbon attacks the cyclic ketone carbonyl of E, forming a new C–CH2NO2 bond while the C=O pi pair moves onto oxygen. protonation by triethylammonium or another proton donor gives the tertiary alcohol F and regenerates base. This is a Henry reaction, also called a nitroaldol reaction: the former ketone carbon now bears both OH and CH2NO2.
The nitro group remains attached to its original nitromethane carbon; attack through oxygen would give a different connectivity. The methyl acetal and the original alcohol substituents need not be cleaved or displaced. This addition produces the isolated connectivity without requiring dehydration to a nitroalkene. The stated complex mixture is compatible with competing additions or stereoisomers, but the scheme supplies no basis for claiming F is the unique or kinetically dominant product. Only its formation is being explained.
water achieves substantial binding through a localized O–H···O hydrogen bond; benzene achieves a comparable total through a larger set of weaker contacts. In the water dimer, one water is the hydrogen-bond donor and the other presents an oxygen lone pair region as acceptor. The interaction includes electrostatics, induction interaction, dispersion and short-range exchange repulsion; a hydrogen bond is not a separate fundamental force that can simply be added to all those terms again.
For benzene, draw a T-shaped edge-to-face pair or a slipped parallel pair. Its larger, more polarizable electron clouds provide appreciable London dispersion. Its quadrupolar charge distribution also makes the energy orientation dependent, despite each molecule having no permanent dipole by symmetry. An edge C–H approaching the other ring's pi region is therefore preferable to a model of two structureless, noninteracting dipole-free objects. Dispersion attracts while Pauli repulsion limits close overlap. Several distributed contacts can sum to the scale of one water–water hydrogen bond; counting only the number of conventional hydrogen bonds would miss them.
The comparison concerns the given gas-phase pair interaction energies, not an equality of every solvent-dependent association free energy. Correlated primary calculations explicitly identify both T-shaped and parallel-displaced benzene-dimer minima, rather than a uniquely preferred perfectly eclipsed sandwich.[ref]
Estimates of the Ab Initio Limit for π–π Interactions: The Benzene Dimer by Mutasem Omar Sinnokrot and others. 2002, abstract and pp. 10887–10893, comparison of the three dimer geometries
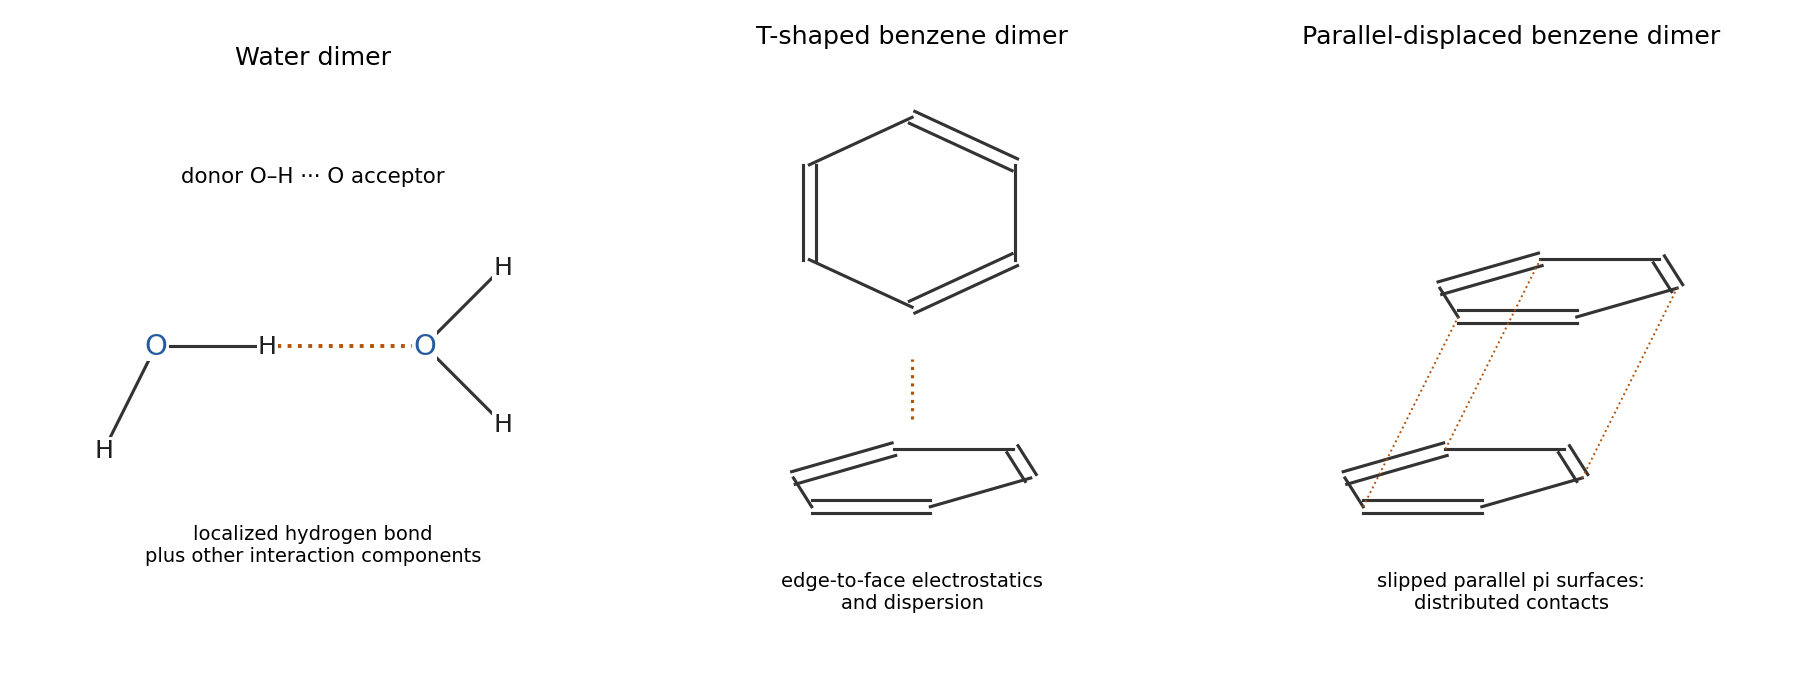
Original schematic geometries of a hydrogen-bonded water dimer and T-shaped and parallel-displaced benzene dimers
. Details. 
Association in solution includes replacement of solute–solvent contacts, not just formation of the isolated dimer. cyclohexane solvates the benzene surface well through dispersion. Much of the London dispersion gained on bringing two benzene molecules together replaces comparable benzene–cyclohexane contacts, leaving only a modest net gain before the translational and orientational entropy penalty. cyclohexane cannot provide the complementary donor/acceptor interactions available when two waters hydrogen-bond, so water association gains a more distinctive interaction relative to its separated solvated monomers.
Using the same concentration standard state, and . For illustration at 298 K and , the ratio of the two supplied association constants corresponds toThus the water-molecule pair can associate more favourably in this solvent even though the stipulated isolated-pair binding energies are similar. Gas-phase potential energy alone omits both differential solvation and the entropy of association.
In water, separated benzene molecules require separate hydration environments around nonpolar surface. Bringing them together buries part of that surface and reorganizes the surrounding water. The resulting solvent contribution favours association relative to cyclohexane, where benzene was already well solvated by dispersion. This solvent-mediated contribution is the hydrophobic effect, not formation of a new covalent bond or a claim that benzene has become a conventional hydrogen-bond acceptor like oxygen.
The supplied constants imply a thirtyfold enhancement. With matched standard states,at an illustrative 298 K. The water value is still slightly below , so enhancement relative to cyclohexane does not mean a negative standard association free energy for a 1 M standard state, nor does it imply that dilute benzene is all dimer. No universal “entropy only” explanation is needed: solvent enthalpy and entropy both contribute and depend on temperature.
TODO: user-deferred biology subquestion. This part is intentionally unsolved under the instruction to skip biology. No biological mechanism, sequence analysis or protein-folding research has been undertaken.
TODO: user-deferred biology subquestion. This part is intentionally unsolved under the instruction to skip biology. No biological mechanism, sequence analysis or protein-folding research has been undertaken.
TODO: user-deferred biology subquestion. This part is intentionally unsolved under the instruction to skip biology. No biological mechanism, sequence analysis or protein-folding research has been undertaken.
TODO: user-deferred biology subquestion. This part is intentionally unsolved under the instruction to skip biology. No biological mechanism, sequence analysis or protein-folding research has been undertaken.
Articles by others on the same topic
There are currently no matching articles.