The 2021 paper occupies bundle pages 74–78; its cover establishes the year despite the attachment's 2020 filename.
Determining antiferromagnetic magnetic order is a good reason to use neutron scattering. The neutron has a magnetic dipole moment, so its scattering amplitude includes the spatial arrangement of electronic magnetic moments. Alternating magnetic moments can produce magnetic neutron diffraction peaks even when there is no change of the nuclear crystal structure. Ordinary nonresonant X-ray diffraction primarily measures the electron density, and is much less sensitive to this magnetic order.
The neutron is nonrelativistic at this kinetic energy. Its de Broglie wavelength isIn elastic scattering, Bragg's law gives . The measured total deflection is , so it must be halved before taking the sine. The reflection order is absorbed into the Miller indices of the diffracting lattice planes. In the order of the four measured rings, the interplanar spacings areThe last digits give the numerical calculation, rather than accuracy beyond the supplied angular precision.
For the cubic crystal system, the Miller indices satisfyThe inverse squared interplanar spacings have ratios approximately . With the lattice constant close to , the appropriate squared-index sums are . Thus the four powder diffraction rings areThe braces collect all symmetry-related lattice planes with the same interplanar spacing. In particular, is forbidden for both a body-centered cubic lattice and a face-centered cubic lattice, but occurs for a primitive cubic lattice. These observations identify the Bravais lattice as primitive cubic lattice, with a multi-atom atomic basis, consistent with a perovskite structure.
Each ring independently gives :Their average is . Allowing for the rounded measured angles, a suitable final value isA larger centred unit cell could relabel reflections, but would conflict with the stated approximate lattice constant.
An applied magnetic field changes the alignment of the manganese magnetic moments. The component of their magnetic structure factor perpendicular to the scattering vector contributes to magnetic neutron diffraction. A field-induced uniform magnetization has the same translational periodicity as the nuclear crystal structure, so its scattering amplitude can occur at the existing reciprocal lattice vectors. The diffraction wave intensities can therefore change without moving the peaks. Their angular positions still measure the same interplanar spacings. This mechanism does not require a measurable change of the lattice constant.
A consistent mechanism is a transition to G-type antiferromagnetism at the Néel temperature. On the manganese primitive cubic lattice, take the magnetic moments to beEvery nearest neighbour has the opposite magnetic moment. The nuclear crystal structure can retain its original lattice constant, while the magnetic order introduces the propagation wavevectorSumming the scattering amplitude from the magnetic moments over sites gives peaks at , where is a nuclear reciprocal lattice vector. Thus all three components of the new Miller indices, expressed relative to the original unit cell, are half-integers. The two smallest squared magnitudes in units of areThe first predicts , agreeing with the new ring. The next squared magnitude is , outside the measured angular range. Taking the observed first magnetic ring as the calibration, Bragg's law predictsUsing the independently fitted lattice constant gives , within the input rounding. One ring alone does not uniquely determine a magnetic order; this proposed nearest-neighbour antiferromagnetic pattern explains both the observed ring and the count of two new rings in the specified interval.
The Bravais lattice is a face-centered cubic lattice. Four inequivalent representatives of its lattice points in the conventional cubic unit cell, in fractional crystal coordinates, areBoundary copies in the drawing do not constitute additional independent lattice points. Attach the two-atom atomic basisto each lattice point. The resulting aluminium phosphide crystal structure is the zinc blende structure. For example, the four phosphorus positions reduced into the conventional unit cell are , , , and , reproducing the heights in the source drawing. There are four aluminium and four phosphorus chemical atoms per conventional unit cell, but only two chemical atoms per primitive unit cell.
Given independent primitive lattice translation vectors , defineThen . The reciprocal lattice consists ofequivalently all wavevectors for which for every direct lattice vector .
For the face-centered cubic lattice, chooseTheir scalar triple product is , and direct evaluation of the cross products givesEvery integer combination has coordinates with all even or all odd. Conversely, for such coordinates the coefficients are integers, so every such point occurs. The even-coordinate points are the corners of cubes of edge , and the odd-coordinate points are their body centres. ThereforeThis is a property of the Bravais lattice; the two-atom atomic basis changes structure factors rather than these reciprocal lattice translation vectors.
There are two chemical atoms in the primitive unit cell, hence six displacement coordinates and six phonon branches at each wavevector. Three rigid translations cost no restoring potential energy in the long-wavelength limit. ConsequentlyFor propagation along , one acoustic phonon has longitudinal polarization along and two have transverse polarization along . The cubic crystal symmetry makes those two transverse directions equivalent. In terms of the elastic stiffness tensor and the mass density,Thus there are two distinct acoustic velocities in general, one longitudinal and a doubly degenerate transverse value. This degeneracy of normal modes concerns the two transverse polarizations; it does not imply equality with the longitudinal speed of sound.
Subtracting the three acoustic phonon branches from the six displacement normal modes leavesAt small wavevector, these involve relative motion of the two atomic basis sites rather than a rigid translation of the primitive unit cell. Their angular frequencies remain nonzero at the zone centre. The count uses the primitive unit cell; using all eight chemical atoms in the conventional unit cell would spuriously multiply the number of distinct branches in the first Brillouin zone.
Use the long-wavelength linear dispersion relation as an estimate. The phonon energy is thenThis extrapolates the measured low-wavevector speed of sound to the point, the first Brillouin zone boundary along . It is not an exact phonon dispersion relation there: the discrete crystal structure generally bends the branch, and the initial slope alone cannot determine its boundary energy. In particular, is not a reciprocal lattice vector of the face-centered cubic lattice; the smallest such axial vector is . There is therefore no reason for this acoustic phonon energy to vanish.
In the diamond cubic structure of silicon, the longitudinal acoustic phonon branch and longitudinal optical phonon branch meet at . The two atomic basis sites are identical, and the associated crystal symmetry protects their boundary degeneracy of normal modes. In aluminium phosphide, the two sites contain different chemical atoms. The corresponding zinc blende structure lacks the site-exchange symmetry, and these two longitudinal branches generally split at .
A simple way to see the splitting is a diatomic-chain dispersion relation with equal springs, representing successive atomic planes by alternating masses at spacing . With effective spring constant , its angular frequencies obeyAt , , so the inter-sublattice coupling cancels and the two squared angular frequencies are and . Equal masses give a contact; unequal masses giveIn aluminium phosphide, the heavier phosphorus plane moves in the lower-frequency edge mode, and the lighter aluminium plane in the higher-frequency mode in this model. Their similar atomic masses explain why the splitting is relatively small. Different electronic environments can also change the force constant matrix. The chain calculation illustrates the mechanism; a quantitative three-dimensional phonon spectrum requires its full dynamical matrix. It does not assert that all transverse acoustic and optical branches meet in silicon.
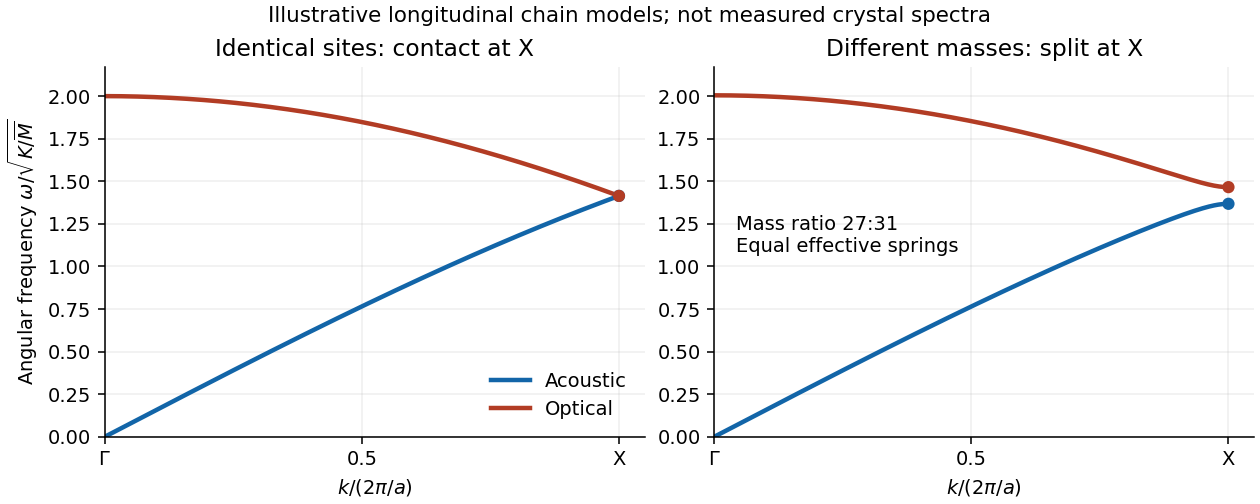
Equal-mass and alternating-mass chain models illustrating longitudinal branch contact and splitting at X
. Details. 
In the harmonic approximation, each in-plane displacement coordinate contributes one quadratic kinetic energy term and one quadratic restoring potential energy term. The equipartition theorem assigns to each. There are two such coordinates per chemical atom, henceHere the fixed-size heat capacity is at constant area in this two-dimensional model. On cooling, phonon excitation freezes out and the heat capacity decreases. For each nonzero angular frequency, the quantum harmonic oscillator contributeswhich is below at finite and approaches zero as . Two linearly dispersing in-plane acoustic phonon branches give the familiar two-dimensional Debye model scaling at sufficiently low temperature. Out-of-plane modes are excluded by the specified model.
For a nondegenerate two-dimensional electron gas, the two quadratic momentum terms supply each by the equipartition theorem. There is no lattice restoring potential energy for a free electron. ThusOn cooling below the Fermi temperature, the Pauli exclusion principle restricts the thermally active electrons to within approximately of the Fermi energy. The linear low-temperature heat capacity of a two-dimensional Fermi gas isThe heat capacity therefore decreases from its classical value as quantum degeneracy becomes important. The particle number of electrons and sample area are held fixed when differentiating the internal energy.
Let the sample area be , set the band touching energy to zero, and write . Assume one electronic valley in each energy band with two spin states. The near-edge parabolic dispersions areUnder periodic boundary conditions, each spin occupies reciprocal-space area . Counting the quantum states inside a circle givesIts derivative is the constant two-dimensional free-electron density of states. Defining , the signed-energy density of states isThe total electronic density of states is therefore on either side of zero, not at every signed energy. If instead one counts positive-energy excitations, the electron and hole species each have density of states , giving when summed. A common additional valley degeneracy multiplies all these quantities by that factor. The effective mass approximation assumes thermally sampled energies remain within the near-edge regions of both energy bands.
Equal effective masses and equal spin and valley degeneracies give electron-hole symmetry. In the intrinsic semiconductor, charge neutrality requires equal numbers of conduction band electrons and valence band holes. It therefore pins the chemical potential to the band touching point, . The Fermi-Dirac distribution givesMeasured relative to the filled valence band at zero temperature, the thermal internal energy includes both species. Removing a valence band electron at costs the positive hole energy , just as adding a conduction band electron at does. HenceDifferentiate at fixed area and fixed total electronic particle number, allowing creation of electron-hole pairs:There is no conflict with the equipartition theorem. The normalization counts only the excited conduction band electrons, while the heat capacity includes both electrons and holes; moreover, is not fixed. In fact , and differentiating supplies another equal contribution to the heat capacity. Finally, gives occupation at the touching point, so the excitation gas is not in the classical dilute limit even though exceeds the vanishing band gap.
For the degenerate electron gas, let include both spin states. The Sommerfeld expansion givesFor the gapless symmetric intrinsic semiconductor, part (d) gave the same coefficient with . Thus for equal sample areas and equal spin and valley degeneracies,The equality often intended in this comparison requires the same density of states. The source sets but does not explicitly equate them to the electron gas mass . Without that additional assumption the ratio is , or more generally if sample areas or degeneracy factors differ. For example, with equal areas satisfies the stated equal-effective mass condition in the semiconductor and gives ratio two. The filled Fermi sea does not add a factor : only the thermal shell near its Fermi energy contributes to its heat capacity.
Undoped gallium arsenide is expected to be diamagnetic at zero temperature. Its valence band is filled and its conduction band empty, so there are no mobile conduction band electrons to give Pauli paramagnetism. The specified absence of localized magnetic moments removes a Curie law response and the usual mechanism for ferromagnetic ordering. The remaining induced orbital response is diamagnetism. This classification uses the ordinary weak-field band insulator picture; the band gap suppresses thermally generated carriers as .
The number of formula units per volume follows from the mass density:Each neutral formula unit contains electrons, so . Let be the electron-weighted mean-square orbital radius, averaged over all these electrons. The supplied Larmor-Langevin susceptibility then gives the dimensionless SI volume magnetic susceptibilitywhere is the Bohr radius. An explicit rough atomic-scale assumption gives .
The quoted bulk data do not determine : a nuclear charge does not specify an orbital radius in a screened many-electron chemical atom. Consequently the numerical estimate just made has atomic-model uncertainty; it is not a deduction with guaranteed 30 percent accuracy. For example, the alternative assumed gives four times the magnitude while leaving every supplied bulk number unchanged. Atomic wavefunctions, an explicit screened-orbital model, or measured atomic magnetic susceptibilities of materials exhibiting diamagnetism would be needed to support that accuracy. Keeping explicit makes the later comparison independent of an unadvertised radius assumption. The bare electron mass belongs in this atomic formula; the conduction band effective mass is relevant instead to the large donor envelope in part (c).
In the hydrogenic donor state approximation, the screened Coulomb potential energy is and the kinetic term uses . Rescaling the hydrogen atom gives the effective Bohr radius of a hydrogenic donor and the positive binding energyThereforeThe radius here is the exponential length of the ground-state wavefunction, not its root-mean-square radius. With , radial integration gives and .
For dilute independent donors of number density , a bound spin one-half electron has magnetic moment magnitude , with the Bohr magneton defined using the bare electron mass. Its weak-field Curie law response isFor the orbital response, expand the effective mass Hamiltonian in a uniform magnetic field, using . The spherical ground state has zero orbital angular momentum; the quadratic term raises its energy byDifferentiating this energy twice and using givesThe donor-only magnetic susceptibility cancels whenThe negative sign of drops out of the linear magnetic susceptibility. This crossover is far below , consistent with donor binding. It assumes sufficiently weak magnetic field, negligible exchange between donors and a thermal scale above any inter-donor splitting; sufficiently dilute donors can satisfy those assumptions. The background diamagnetism from part (b) is excluded from this cancellation.
A hydrogenic donor state has an exponentially decaying envelope . Neighbouring donors at typical separation therefore have exponentially small orbital overlap. The corresponding hopping integral is of order , apart from algebraic factors and factors from the number of neighbours. Hybridizing many such localized states produces an impurity band with a bandwidth of the same scale:The order-one constant absorbs details of the donor spacing and envelope. The exponential dependence expresses quantum tunnelling between weakly overlapping donor orbitals, rather than thermal excitation over the donor binding energy.
Use the stipulated noninteracting impurity band approximation, with one donated electron per donor and two available spin states per donor orbital. The energy band is half filled. Approximating its spin-summed density of states per volume near the Fermi energy by gives the zero-temperature Pauli paramagnetismWith ,A factor of order one depends on the density of states shape; using the conduction band -factor rather than silently setting matters here.
To locate the comparison with the magnitude of the undoped Larmor-Langevin susceptibility, put . Numerically,For the stated atomic-scale estimate in part (b), equality with requiresThus the dilute-branch crossover iswell below . Within this model the positive spin response exceeds the negative background magnitude for lower dopant densities on this dilute branch: the exponential narrowing of wins over the loss of donors. Indeed for . This is not a claim of ferromagnetism.
The degenerate Pauli paramagnetism calculation requires , with an unimportant order-one filling factor. At the estimated crossing,The weak-field condition also requires . For a different assumed atomic , use in the displayed equation for . An order-of-magnitude background gives the same density order and a crossover scale of order . At higher temperature than this extremely narrow bandwidth permits, the enhancement cannot be used; the independent-donor Curie law is the appropriate thermal scale if interactions remain negligible. At very small dopant density, disorder, localization and Coulomb interactions can also invalidate the assumed noninteracting impurity band. The formal divergence as is therefore a limit of the stated zero-temperature band model, not a prediction for isolated donors at a fixed nonzero temperature.
Articles by others on the same topic
There are currently no matching articles.