Molecular vibration changes internal separations and bond angles, unlike the fixed internal molecular geometry of a rigid rotor. Its coupling to molecular rotation makes measured rotational constants depend on the vibrational quantum state.
A diatomic molecule has one internal stretching coordinate: its two atomic nuclei approach and separate about an equilibrium bond length. The diatomic bond stretch turns this motion into a particularly useful model. The local potential energy curvature supplies the restoring force, while the reduced mass accounts for both moving partners. Thus the same chemical bond can vibrate more slowly after an isotope substitution even when its electronic structure is nearly unchanged.
This separation makes infrared spectroscopy informative: transition frequencies constrain chemical bond stiffness, and isotope scaling of diatomic vibration tests the mass dependence. The University of Florida HCl and DCl FTIR experiment illustrates the measurement. Detectability also requires vibrational infrared activity; having vibrational energy eigenvalues alone does not guarantee an infrared absorption line.
Loss of excess vibrational excitation to other molecular degrees of freedom or to the surroundings. It can occur within one electronic state without photon emission, reducing the energy available to subsequent fluorescence.
A mass-weighted displacement coordinate along an eigenvector of the molecular harmonic force-constant matrix after rigid translations and rotations are removed. An orthonormal normal-mode basis gives kinetic energy and harmonic potential . This use of normal coordinate concerns molecular vibrations, rather than normal coordinates of a spacetime metric.
In the quadratic vibrational term expression , the constant is dimensionless, while has units of spectroscopic wavenumber. A positive value makes adjacent level spacings decrease. For an exact Morse oscillator, . Using only two low-lying transitions to extrapolate this identity to a real molecule assumes the higher-order anharmonic terms are negligible all the way to dissociation.
Coordinates belonging to inequivalent irreducible representations cannot have an invariant bilinear cross term. Two copies of the same real irreducible representation can: their componentwise dot product is invariant. Diagonalizing the multiplicity-space force-constant matrix produces the normal modes. Symmetry alone therefore does not require two coordinate sets to have zero coupling before that diagonalization.
Coupling between electronic and nuclear vibrational motion. Dependence of an electronic electric dipole transition moment on a normal mode coordinate can give intensity to a transition whose purely electronic matrix element vanishes at the equilibrium geometry.
An otherwise forbidden electronic transition gains intensity when accompanying molecular vibration supplies a symmetry factor that permits a nonzero electric dipole transition moment. A symmetry-allowed total vibronic state specifies a possible transition, while the coupling determines its actual strength.
A joint electronic and vibrational quantum state of a molecule. In a product approximation its irreducible representation is the tensor product of group representations of the electronic and vibrational factors.
The angular momentum associated with motion in a degenerate bending normal mode of a linear molecule. A one-quantum bend has , and the total rotational quantum number obeys .
Spectroscopy of simultaneous changes of vibrational and rotational state. The transition wavenumbers combine vibrational energy differences with rotational constant terms. The allowed rotational changes depend on the molecule and the transition type.
A symmetric-top vibrational band whose transition dipole is perpendicular to the symmetry axis, with . Unresolved Q subbranches can create a regular sequence controlled mainly by .
A symmetric-top vibrational band whose transition dipole is parallel to the symmetry axis, with in the elementary model.
The rovibrational spectroscopy branch in which absorption raises the total rotational angular momentum quantum number by one. A label conventionally specifies the lower-state value .
The rovibrational spectroscopy branch with unchanged rotational angular momentum quantum number. Differences between upper and lower rotational constants can resolve its transitions into closely spaced lines. Allowed lower-state values depend on the angular momenta of both states.
The rovibrational spectroscopy branch in which absorption lowers the total rotational angular momentum quantum number by one. A label conventionally specifies the lower-state value .
The observed collection of lines or bands from vibration-rotation transitions. In the usual parallel diatomic infrared band, P and R branches have and the Q branch is absent.
A transition starting from a thermally populated vibrationally excited state. Its intensity depends on the lower-state population through the Boltzmann distribution. Anharmonicity and coupling between normal modes can shift it away from the corresponding vibrational fundamental.
Excitation of one quantum of a normal mode from the vibrational ground state. For an infrared-active mode in the linear-dipole harmonic approximation, this is the allowed absorption underlying the usual fundamental band. A vibrational overtone excites two or more quanta from the ground state.
In the independent harmonic approximation, each diatomic molecule has one bond-stretching quantum harmonic oscillator with energies . Write . Its canonical partition function is , and differentiating its logarithm givesFor , excitation is exponentially suppressed and the heat capacity tends to zero. At high temperature it tends to , one contribution from each quadratic kinetic and potential term. The zero-point energy does not contribute to the derivative with temperature. For a dilute gas whose rotations are already active, adding translation and rotation gives molar before vibration is activated and afterwards, provided dissociation and electronic excitations are negligible.
Let be the instantaneous bond length, its equilibrium value, and the stretching displacement. Assume one fixed electronic quantum state, a smooth potential energy with a nondegenerate local minimum at , and omit molecular rotation initially. The Taylor series has no linear term because :Keeping only the quadratic term is the harmonic approximation near a potential minimum. The bond force constant has units . It measures local stiffness; it is not the bond-dissociation energy, which concerns the depth and distant shape of the entire well.
The two-particle relative coordinate explains the reduced mass. For stretching along a fixed axis in a stationary centre of mass frame, let be the nuclear displacements from equilibrium. Then and , soHere is the internal kinetic energy, and are the two nuclear masses. The lighter partner moves farther; the reduced mass is smaller than either individual mass. The restoring force gives simple harmonic motion:The angular frequency is , the ordinary frequency is , and is the corresponding spectroscopic wavenumber; is the speed of light. Use in to obtain in . The symbol here is a bond force constant, not an angular wavenumber.
For an original dimensionless example, choose and in consistent model units. Then and . Releasing at with gives and , . Each atomic nucleus moves only half the stretching amplitude, while the conserved internal mechanical energy above is . Fixing one partner instead would give , demonstrating why a one-body calculation changes the model.
The bond force constant is the curvature of the effective potential energy at the equilibrium bond length. For a diatomic bond stretch, it plays the role of a spring constant. Its definition is local: two wells with the same curvature can have different bond-dissociation energies.
Quantizing the stretching coordinate gives a quantum harmonic oscillator with reduced mass :Here is the vibrational quantum number and the reduced Planck constant. Consecutive energy eigenvalues have gap ; the ground state has zero-point energy above the well minimum. The offset cancels from every transition energy.
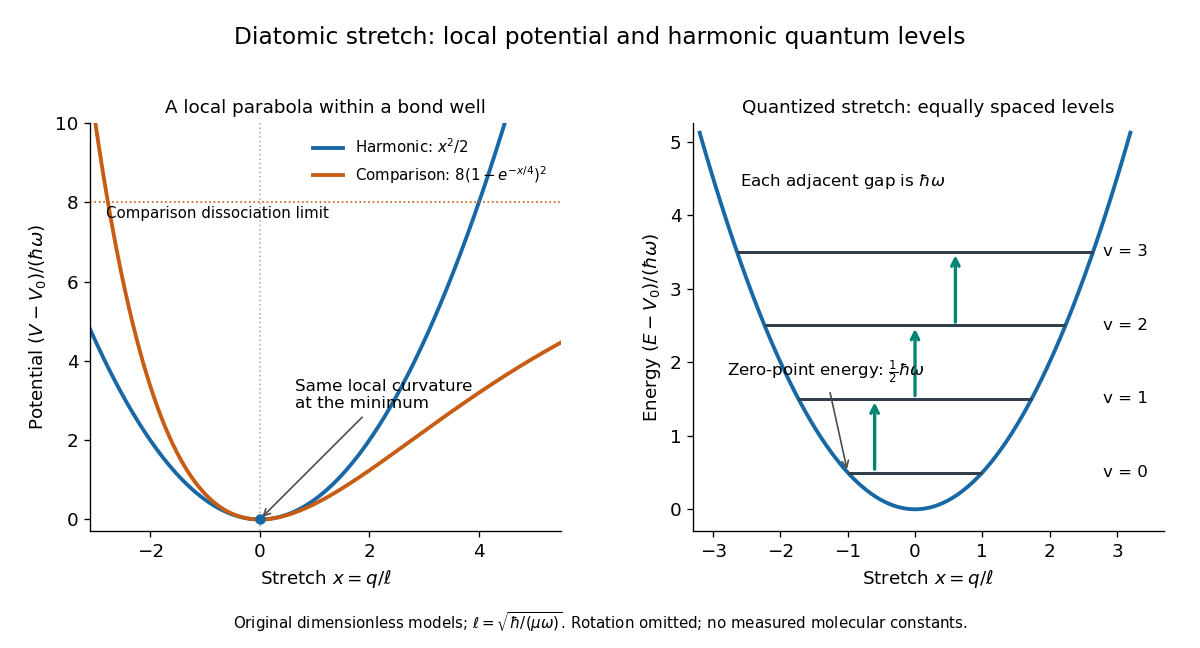
Local harmonic approximation near a potential minimum and equally spaced vibrational energy eigenvalues
. Original dimensionless model with and . The blue parabola is ; levels are . The orange comparison is the stipulated Morse potential , with the same curvature at ; it is not a fit to hydrogen chloride. Arrows show adjacent vibrational gaps, with molecular rotation omitted. Details.A short proof uses creation and annihilation operators: with , write . The Quantum Hamiltonian becomes . Nonnegativity of gives the lowest level; repeated application of raises its energy by each time. This also gives andThe ground state therefore has a nonzero spread of bond lengths, even though its mean stretching displacement vanishes. A stationary energy eigenstate has a time-independent position probability density; the classical oscillation curve is not a trajectory of an atomic nucleus in that state.
The parabolic model formally extends to the whole real line, whereas an actual bond length satisfies . It is useful for low levels concentrated near , with typical stretching displacements much smaller than . It fails near breakup of the chemical bond and when neglected anharmonicity becomes substantial.
Keep the same bond force constant and approximate the nuclear masses of , , and by , , and , respectively. Here is the unified atomic mass unit, and denotes deuterium. These are chosen mass number approximations, not measured atomic masses or natural-abundance averages. With the same chlorine isotope in both molecules,This is the isotope scaling of diatomic vibration prediction. Doubling the light partner's mass nearly doubles the reduced mass, lowering the frequency by about . The factor differs from because the chlorine partner also moves.
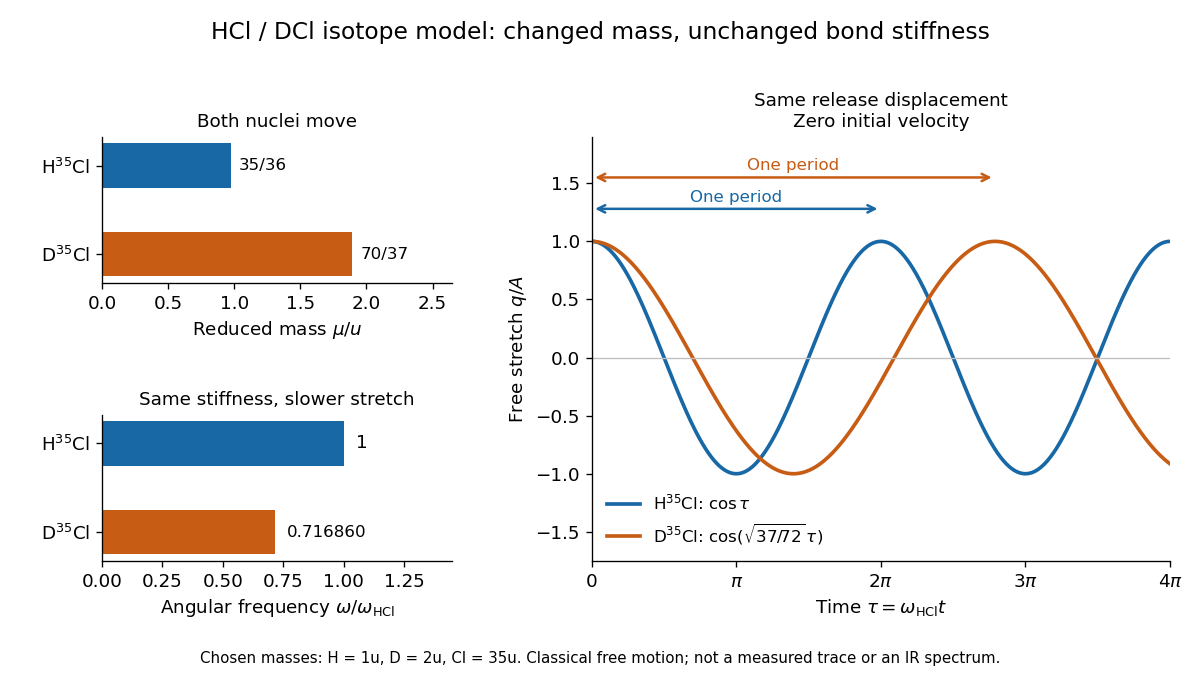
For the same small initial stretch and zero velocity, both models start with the same internal mechanical energy , but the DCl oscillation period is longer by . In the centre of mass frame, and for HCl; and for DCl. These ratios explain which partner moves and are separate from the change in frequency.
The common- assumption belongs to the leading Born-Oppenheimer approximation: fixed nuclear positions and electric charges determine the electronic potential energy, while nuclear masses govern motion on that curve. Quantitative comparison requires actual atomic masses, departures from that approximation, and separation of anharmonicity and molecular rotation from the measured band. Even with common , the quantum harmonic oscillator predicts a different zero-point energy for each isotope.[ref]
In the ideal electric-dipole interaction, a vibrational transition needs a nonzero matrix element of the electric dipole moment. For a diatomic bond stretch, expand the body-axis dipole component as . A changing electric dipole moment, rather than a permanent value alone, permits the harmonic fundamental. Keeping only the linear term and harmonic wavefunctions gives .[ref]
14: Harmonic Oscillators and IR Spectroscopy, UCD Chem 110A by Delmar S. Larsen. Lecture 14, introductory recap and infrared-activity section: dipole change and adjacent-level selection
The derivation is sharper than a picture of a shaking molecule. Orthogonality removes the constant term for , and creation and annihilation operators giveHere is the Kronecker delta. Thus absorption raises by one when the dipole derivative and angular factor are nonzero, and its photon satisfiesA nonzero vibrational matrix element is only one factor: rotational electric-dipole selection rules, initial populations, and spectral resolution also constrain visibility.
For an isolated diatomic molecule with identical atomic nuclei, such as molecular hydrogen in its symmetric electronic ground state, stretching preserves zero electric dipole moment. By symmetry, this homonuclear stretch is infrared inactive in the ideal dipole model, although it still vibrates. Other interactions or environments can permit detection.[ref]
14: Harmonic Oscillators and IR Spectroscopy, UCD Chem 110A by Delmar S. Larsen. Lecture 14, introductory recap: dipole-change criterion; the identical-nuclei case follows here by symmetry
The University of Florida's Rotation Vibration Spectrum of the HCl Molecule, University of Florida laboratory protocol, printed pp. 2–3 and 10–11, uses a scanning Michelson interferometer for Fourier transform spectroscopy. An air-filled silicon-window cell supplies the background. Supervised preparation adds concentrated hydrogen chloride liquid; the cleaned cell subsequently receives DCl liquid. Acquisition settings are identical.
The readout is transmission or absorbance against spectroscopic wavenumber, resolving molecular rotation alongside stretching. Background addresses instrument and cell contributions; cleaning addresses carryover. Excessive absorption impairs analysis. Assigning isotopes and rotational branches precedes fitting band parameters to test isotope scaling of diatomic vibration. These are prescribed measurements, not numerical observations supplied here.[ref]
Rotation Vibration Spectrum of the HCl Molecule, University of Florida laboratory. printed pp. 10–12: Measurements, Preliminary Analysis and Data Analysis, including matched scans, excessive absorption, isotope assignments and fitted band origin
For a transparent ideal model, add the linear rigid rotor to the harmonic vibrational energy levels and assume the same frequency-valued rotational constant in both vibrational states:For a spin-free diatomic molecule with no electronic angular momentum, electric-dipole selection rules give for this vibrational absorption. Subtracting initial from final energy eigenvalues yieldsThese are two rotational branches about the vibrational band origin, not a single stretch line. For stipulated dimensionless values and , the first three R-branch positions are , and the first three P-branch positions are . There is no transition. The numbers illustrate level subtraction; they are not HCl measurements. Real upper and lower rotational constants differ, and centrifugal distortion shifts lines further.
A vibrational overtone excites more than one vibrational quantum: is the first overtone, and the second. Such transitions vanish in the combined harmonic-potential and linear-dipole approximation. Anharmonicity changes wavefunctions and level spacing; nonlinear terms in the electric dipole moment can also supply otherwise missing transition matrix elements.[ref]
14: Harmonic Oscillators and IR Spectroscopy, UCD Chem 110A by Delmar S. Larsen. Lecture 14, overtone section, Table 1 and following potential-expansion discussion
For an original illustrative level model, setThen and . With chosen , the fundamental gap is and the first-overtone gap , below twice the fundamental, . This low-level illustration does not determine transition intensity or justify extrapolation to arbitrarily large . It shows why even a well-resolved fundamental band origin need not equal the harmonic curvature scale .
Articles by others on the same topic



Molecular vibration refers to the oscillatory motion of atoms within a molecule around their equilibrium positions. This phenomenon occurs because molecular bonds can be thought of as springs that can stretch and compress, allowing the atoms to move closer together or further apart. During vibration, different types of motions can occur, including: 1. **Stretching**: This can be further divided into: - **Symmetric stretching**: Both bonds are elongated or shortened simultaneously.