Molecular rotation changes a molecule's orientation. Its rotational kinetic energy depends on the arrangement and masses of its chemical atoms. A rigid rotor freezes internal distances to isolate this motion; rotational spectroscopy tests the resulting energy eigenvalues through transitions between quantum states. The molecular beam electric resonance experiment provides a complementary probe of the coupling to an electric field.
The dependence of rotational energy on the vibrational state. For a diatomic molecule a common leading approximation is , so vibrational satellites can have different rotational line spacings.
The first-order dependence of a polyatomic rotational constant on vibrational occupations , where is the degeneracy of normal mode . The factor accounts for all of that mode's zero-point energy. A single band's change in rotational constant generally determines one , rather than every zero-point correction needed for .
A rigid rotor models rotational motion with fixed internal separations. Its use in the synthetic Stark effect figure omits vibrational motion and additional interactions in real molecules.
Rigidity fixes the molecular geometry, not the orientation, and does not require a spherical mass distribution. Remove motion of the centre of mass, choose principal axes of the inertia tensor, and write the classical rotational kinetic energy asHere are the principal moments of inertia, components of angular velocity, and components of angular momentum. Quantization replaces these last components by angular momentum operators. Different principal moments of inertia produce different spectra: the single-constant linear rigid rotor is a special case, not a formula for every molecule.[ref]
Lecture 24 Supplement: Energy levels of a Rigid Rotor by Dudley Herschbach. printed p. 1: principal-axis energy and classification; pp. 4–5: asymmetric rotors
For a rigid heteronuclear diatomic rotor, and electric-dipole absorption has . Therefore the line frequency is and adjacent absorption lines are spaced by . With reduced mass , the bond-length estimate is . The conversion assumes frequency rather than angular frequency; centrifugal distortion makes real high-J spacings imperfectly uniform.
A rigid rotor restricted to one angular coordinate . Single-valued spatial states have and energy , with moment of inertia . The zero-angular-momentum level has state degeneracy one; every nonzero has two directions. For labelled molecular ends its high-temperature rotational partition function is , where . Equivalent orientations and nuclear spin statistics require a further molecular counting convention. One quadratic angular momentum gives rotational heat capacity per molecule, rather than for an unrestricted linear rigid rotor.
For a linear rigid rotor with moment of inertia , the energy eigenvalue at angular momentum quantum number is . Each level has statistical weight before additional symmetry or nuclear-spin restrictions.
The angular momentum associated with end-over-end molecular rotation. In an open-shell diatomic molecule it is useful to distinguish from the total , which also contains the electronic spin angular momentum.
The rotational symmetry number corrects a rotational phase-space count for indistinguishable molecular orientations. A linear molecule with identical ends, such as carbon dioxide with identical oxygen isotopes, has ; making its ends isotopically distinct gives . In the high-temperature linear rigid rotor approximation, . Holding the spectrum otherwise fixed, doubling lowers molar rotational entropy by .
For a diatomic molecule, let be the two masses and their fixed separation. The distances from the centre of mass are and . A perpendicular axis through the centre of mass therefore has moment of inertiaThe reduced mass here is not the electric dipole moment, for which we use . For a longer linear molecule, replace by , where is each distance along the axis from the centre of mass. In the point-mass idealization, the axial moment of inertia is zero; turning about that axis changes no nuclear positions, so no independent axial rotational degree of freedom is included.
For the field-free linear rigid rotor, assume a fixed internal quantum state with no electronic angular momentum or electronic spin, and omit couplings to nuclear spin. The rotational Hamiltonian is . Its orientation wavefunctions are spherical harmonics , and its angular momentum eigenstates satisfywhere and . Each belongs to angular momentum eigenstates distinguished by the magnetic quantum number , an instance of degenerate energy levels. The Planck constant is ; is the frequency-valued rotational constant. Notice that the energy eigenvalues scale as , not .
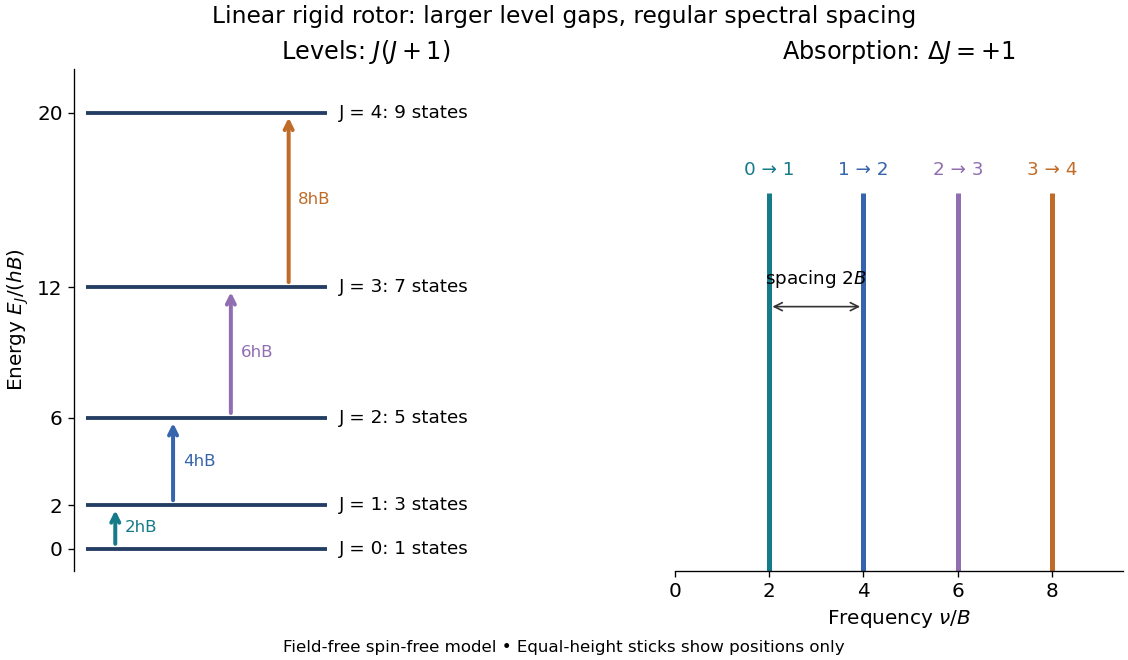
Linear rigid rotor: increasing gaps between energy eigenvalues and equally spaced transition frequencies
. Original dimensionless model: . Upward arrows show photon absorption with ; the sticks show positions only, not measured or predicted intensities. The states at each energy eigenvalue are represented by one horizontal line. Details.
Subtracting consecutive energy eigenvalues givesThus increasing level gaps produce a regularly spaced series of lines at , with spacing . Allowed positions do not guarantee that every line is visible: electric-dipole selection rules, occupation of the initial quantum state, and instrument response also matter.
For a linear rigid rotor, rotational temperature expresses the rotational energy constant in kelvins: , where is the perpendicular moment of inertia. With all integer allowed and degeneracies , its rotational canonical partition function is . It approaches at low temperature and at high temperature. The latter limit gives mean rotational energy and heat capacity per rotor.
In this article the rotational constant is a frequency,Its dimensions are , since the Planck constant has dimensions and a moment of inertia has dimensions . Some authors instead call the energy-valued rotational constant; this is the convention in the earlier synthetic molecular beam electric resonance diagram. A third convention uses the spectroscopic wavenumber , with the speed of light. Always check units before using .
If is known and the two masses are specified, the rigid two-atom model gives the bond length . This is why rotational spectroscopy can constrain molecular geometry. A real fitted constant can depend on molecular vibration, so this inference alone does not establish an equilibrium bond length.
For the field-free, spin-free linear rigid rotor in one fixed internal quantum state, the electric-dipole interaction is . A nonzero permanent electric dipole moment along the molecular axis allows pure rotational photon absorption with and photon emission with . The matrix element between initial and final quantum states must be nonzero: the existence of an energy gap alone is insufficient. Relative to a chosen axis, polarization of an electromagnetic wave along selects ; transverse polarization of an electromagnetic wave permits .[ref]
Lecture 24 Supplement: Energy levels of a Rigid Rotor by Dudley Herschbach. printed pp. 2–3: dipole transition integrals, polarization and selection rules
Hydrogen chloride and carbon monoxide are examples with a permanent electric dipole moment. By symmetry, molecular hydrogen and carbon dioxide have no permanent electric dipole moment in the ideal symmetric internal state. They still rotate and have rotational energy eigenvalues, but the simple permanent-electric dipole spectrum described here is absent. This restriction concerns this interaction and these assumptions, not every possible way of detecting molecular rotation.
Changing an isotope changes a mass and therefore the moment of inertia. If the bond length is held fixed, and every corresponding transition shifts by the same fractional amount. This leading inverse-reduced mass scaling is also used in the more complete diatomic formulation.[ref]
1Σ-Ground State Molecules. §2a, equations (4)–(5): isotope dependence of rotational constants and reduced mass
For a deliberately stipulated carbon monoxide example, take , , , , for both isotopic forms, and . The integer masses are simplifying inputs, not accurate isotopic atomic masses; the shared bond length is an imposed model assumption. Calculation givesIn particular,The heavier form's calculated line is rather than , a decrease of about . These are model predictions from the stated inputs, not published measured frequencies. An isotope changes the spacing as well as the first line; the shift is not a fixed additive offset for the whole spectrum.
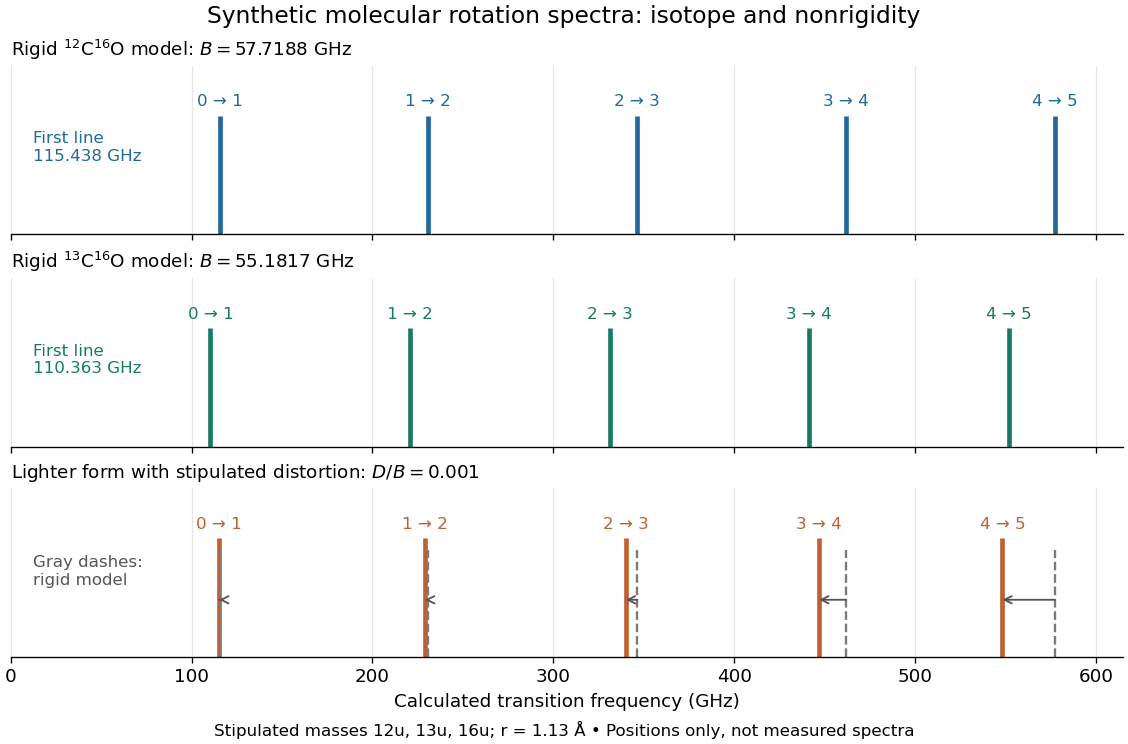
Synthetic rotational spectroscopy comparison: a heavier isotope compresses the rigid spectrum, while centrifugal distortion bends the spacing pattern
. Original equal-height sticks for through , using only the stipulated masses and bond length above. The lower panel additionally stipulates for the lighter form and compares with dashed rigid positions. Neither heights, distortion constant nor line frequencies are measured data; no thermal-population or instrument model is included. Details.
Real chemical bonds can stretch as molecular rotation increases. The resulting larger moment of inertia lowers the energy eigenvalues relative to a rigid rotor. Within one vibrational quantum state, a leading effective correction isBoth and are in frequency units here; labels the vibrational quantum state. For positive , the adjacent-line spacing becomesso the departures grow with . Molecular vibration also changes the effective rotational constant: even the lowest vibrational quantum state does not have motionless atomic nuclei.[ref]
The quartic correction is a limited expansion, not a model to extrapolate to arbitrary . It is small relative to the rigid contribution only while ; eventually higher terms, changed internal motion, or chemical bond breaking matter. In the synthetic plot, the largest upper-state label is , giving a stipulated correction ratio of . No universal maximum valid follows without a particular molecule and precision target.
The quantum numbers also have limits of meaning. Here describes spin-free rotation. In a molecule with electronic spin, rotation alone is commonly labelled , while includes electronic angular momentum; including nuclear spin introduces additional total-angular momentum labels and resolved splittings. An applied electric field can mix different field-free states through the Stark effect. For identical atomic nuclei, the Pauli exclusion principle can restrict combinations of rotational and nuclear-spin states. The simple degeneracy and every-integer- ladder therefore describe the stated idealization, not all observed sublevels.[ref]
Rotational spectroscopy measures transitions between molecular rotation quantum states, usually in low-pressure gaseous molecules. The measured spectral lines become a molecular fingerprint only after rotational spectral line assignment: identify which transitions belong together, determine their quantum numbers, fit a rotational Hamiltonian, and test additional predictions. Line positions constrain moments of inertia, while rotational state populations and transition coupling determine which lines can be detected.
The simplest ordinary microwave absorption ladder belongs to a polar linear molecule obeying pure rotational electric-dipole selection rules. A homonuclear diatomic molecule has no permanent electric dipole moment in its symmetric internal state, but rotational Raman spectroscopy can probe its rotation through anisotropic electric polarizability. The coupling mechanism must therefore be specified before interpreting a missing spectrum.
The existing linear rigid rotor, rotational constant, isotope shift of a linear rotor, and centrifugal distortion explain the forward model. Molecular structure from rotational constants asks the reverse question: which arrangements of chemical atoms could produce the fitted constants? The NIST trace-gas FTMW validation experiment makes the measurement chain concrete and separates identifying a species from measuring its concentration.
A set of spectral lines associated with transitions between molecular rotational energy levels. A polar linear rigid rotor has electric-dipole transitions with ; in the ideal model adjacent absorption frequencies are separated by twice the rotational constant in frequency units.
For a polar rigid symmetric top with its permanent dipole on the symmetry axis, electric-dipole pure rotation has , . The term cancels within each transition; centrifugal distortion can lift the rigid-model coincidence of different K components.
A rotational spectral line assignment associates a measured spectral line with a molecule, an internal quantum state, and an initial and final set of quantum numbers. For the spin-free linear rigid rotor, absorption is labelled . A regularly spaced group suggests a candidate rotational constant, but the lowest line inside a scanned band need not be . Several molecules can contribute to the same spectrum, and an apparent missing member can lie outside the band or below the detection threshold.
First determine line centers and their uncertainties from the instrument's readout, accounting for resolved Doppler shifts or hyperfine structure rather than treating every peak as a separate rotational transition. Then propose integer assignments, fit a rotational Hamiltonian, and predict additional spectral lines. For a weakly distorted linear molecule within one vibrational quantum state, the centrifugal distortion model gives the useful diagnosticHere are all frequencies in . Correct assignments make the second expression a straight line against , with intercept and slope . This follows by division of the transition formula; no additional model of molecular geometry is required. A direct weighted least squares fit of to uses the measured uncertainties. If instead the transformed values are fitted, their uncertainties must also be divided by .[ref]
For an original example, stipulate and , and suppose a scan covers –. The model places four consecutive transitions at , , , and . Their assignments are , hence . An extra stipulated line at is unrelated to this series. All these frequencies are calculated example inputs, not measured OCS data.
At and , the transformed values are and . Their straight line has slope , recovering and . Both intermediate transitions satisfy the same line exactly. Merely relabelling the first visible transition fails this diagnostic. The model predicts a withheld transition at : an additional measurement there would test the assignment rather than merely refit the original four lines.
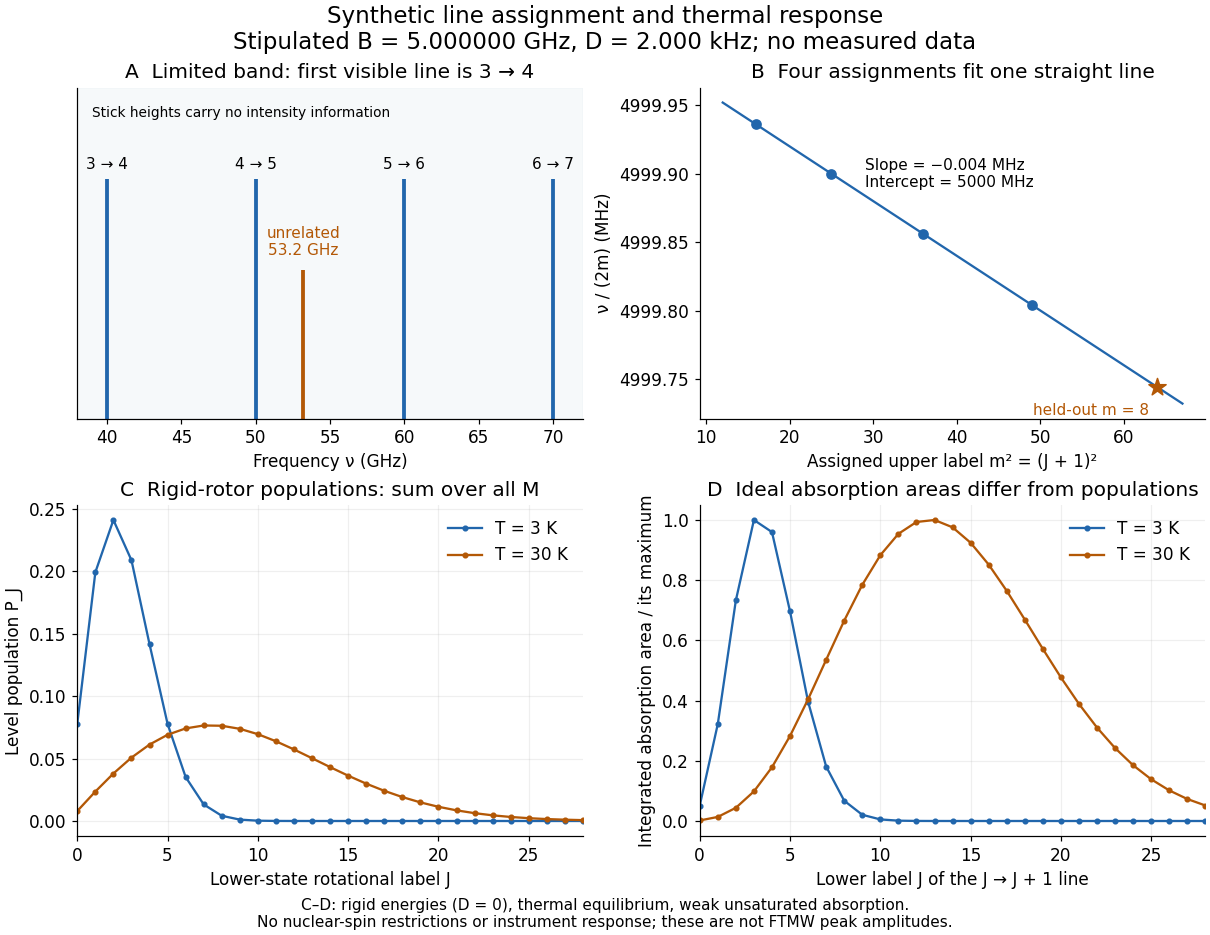
Original synthetic rotational spectral line assignment, fit diagnostic, and rotational state populations
. Stipulated values, not measured data. Upper panels use and ; stick heights show positions only. Lower panels independently use the linear rigid rotor with the same , , and temperatures and . Rotational state populations and ideal integrated absorption areas are different quantities; each absorption curve is normalized separately. Neither is a predicted pulsed FTMW spectroscopy peak amplitude. Details.
An assignment earns confidence when several independently observed spectral lines, including lines outside the fitted interval, agree within uncertainty and expected strong accessible lines are present. Inspect the residual of an inverse problem for each line against : systematic curvature can signal inadequate centrifugal distortion, whereas isolated discrepancies suggest a blend or wrong assignment. Comparing a known sample, a blank, and an isotope shift of a linear rotor supplies different checks. These are general inference controls; the actual reported controls of the NIST trace-gas FTMW validation experiment are described separately. One isolated match or one fitted rotational constant alone cannot certify molecular identity.
For the spin-free linear rigid rotor in thermal equilibrium at temperature , assume all integer are allowed and ignore nuclear-spin restrictions. Each level has quantum states sharing one energy eigenvalue, an instance of degenerate energy levels, so the Boltzmann distribution givesHere is a rotational partition function, the Planck constant, the Boltzmann constant, and a frequency in . Thus is dimensionless. The factor counts magnetic quantum numbers; it does not mean that a whole level is one quantum state. For the stipulated , the most populated levels are at and at .
Population alone is insufficient to predict intensity. In weak, unsaturated absorption, the integrated absorption coefficient includes the transition frequency, the squared electric dipole moment matrix element, and the difference of lower and upper substate populations because stimulated emission returns energy to the field. A pulsed FTMW spectroscopy signal additionally depends on excitation, quantum coherence, cavity tuning, and receiver response.[ref]
Absorption lineshapes by Andrei Tokmakoff. Andrei Tokmakoff, §§6.1–6.2, printed pp. 6-2–6-6, equations (6.8)–(6.10), (6.27)–(6.29): population difference and rotational dipole matrix elements
For an isotropic sample with polarization of an electromagnetic wave along , write . Normalized spherical harmonics giveThe last equality follows from . Combining this sum with the per-substate Boltzmann distribution and subtracting stimulated emission gives relative integrated absorption areasThe common factor includes and cancels upon normalization. This formula describes integrated weak absorption in a thermal, spin-free linear rigid rotor, not arbitrary peak heights. The example's strongest integrated lines originate at and for and , respectively: they differ from the most populated levels. A missing line can reflect population, coupling, coverage, or sensitivity; it does not prove that the corresponding quantum state is absent.
The rotational state populations of a spin-free polar linear rigid rotor are proportional to . Differentiating their logarithm with respect to continuous gives the displayed maximum when it is nonnegative. The actual maximum must be selected among nonnegative integers by comparing adjacent populations. This population maximum generally differs from a maximum in measured spectral line intensity.
Recovering molecular geometry from rotational constants is an inverse problem. In the synthetic assignment, gives . For a diatomic molecule with known masses, rotational constant explains how to infer an effective bond length. For a longer linear molecule, one cannot determine every bond length: several different arrangements can have the same moment of inertia.
For a fixed three-atom linear model with positions and positive masses , subtracting the centre of mass givesThis is one equation for two unknown bond lengths. Even fixing the order of the chemical atoms leaves a family of solutions. Additional isotope substitutions change known masses and supply independent equations, provided the effective molecular geometry can be treated as shared. For nonlinear molecules, three principal moments of inertia give frequency-valued constants , , and ; a single evenly spaced comb is generally insufficient.
Vibrational averaging, unresolved hyperfine structure, internal motion, and centrifugal distortion limit structural inference. A ground-vibrational fitted rotational constant is not automatically the equilibrium constant. Ordinary field-free rotational spectroscopy also gives identical spectra for opposite enantiomers under the usual parity-symmetric molecular Hamiltonian; assigning composition and conformation does not by itself determine handedness. Molecular identification from a rich fingerprint can therefore succeed even when a unique full molecular geometry cannot be reconstructed.
Rotational Raman spectroscopy observes a frequency change of scattered light associated with molecular rotation. The coupling uses the induced electric dipole moment, , where is the electric polarizability tensor. For a linear molecule, the useful anisotropy is : the response differs for an electric field parallel or perpendicular to the molecular axis. A permanent electric dipole moment is not required.[ref]
Absorption lineshapes by Andrei Tokmakoff. Andrei Tokmakoff, §6.2, printed pp. 6-7–6-8, equations (6.40)–(6.44): induced dipole, polarizability tensor and rotational Raman response
For the spin-free linear rigid rotor in one internal quantum state, the anisotropic tensor depends quadratically on the molecular direction. Its angular part has even parity and degree two in spherical harmonics. The triangle inequality for coupled angular momenta permits , and even parity removes odd , leaving . The contribution is unshifted light; shifted rotational lines have . Consequently the rigid-model Stokes shifts, in ordinary frequency units, areSuccessive allowed shifts are separated by , rather than the absorption spacing of pure rotational electric-dipole selection rules. In spectroscopic wavenumber units replace by and by , using the speed of light in for .
A homonuclear diatomic molecule such as molecular hydrogen has no permanent electric dipole moment in its symmetric internal state, so it lacks the ordinary electric-dipole microwave ladder. Its anisotropic electric polarizability can nevertheless give rotational Raman spectroscopy lines. Nuclear-spin symmetry can change the populations of even and odd levels, so the spin-free population example above must not be reused unchanged for molecular hydrogen. The method of coupling, as well as the energy eigenvalues, determines what the experiment can see.
Fourier transform microwave spectroscopy excites gaseous molecules with a pulse of electromagnetic radiation, then records the emitted time-dependent electric field. A Fourier transform extracts transition frequencies from the decaying signal. In a cavity implementation, two facing mirrors form a resonator; tuned excitation, receiver electronics and digitization replace the molecule-counting readout of molecular beam electric resonance. The pulsed radiation prepares quantum coherence between rotational quantum states; it does not set the whole gas spinning at one classical angular velocity.
For documented laboratory context, Evolution of Microwave Spectroscopy at the National Bureau of Standards (NBS) and the National Institute of Standards and Technology (NIST) by F. J. Lovas and others (§5, printed pp. 280–282) describes the NIST cavity instrument, phase-locked sources, mirror tuning and detector-mixer modifications. That account explains one instrument family; it does not establish the electronics of every later commercial spectrometer.
The NIST validation report used carbonyl sulfide in argon to illustrate the readout. A pulsed nozzle injected a molecular beam along a two-mirror cavity. Approximately excitation prepared quantum coherence; a receiver digitized the decaying electric field, then a Fourier transform produced spectral lines. Counterpropagating cavity waves produced a Doppler shift doublet; its midpoint represents one rest-frame transition.[ref]
The Validation of Fourier-Transform Microwave Spectroscopy for Trace-Gas Analysis. The Validation of Fourier-Transform Microwave Spectroscopy for Trace-Gas Analysis, printed pp. 25–26, Figures 2–3: coaxial apparatus, excitation and OCS readout; PDF pages 31–32
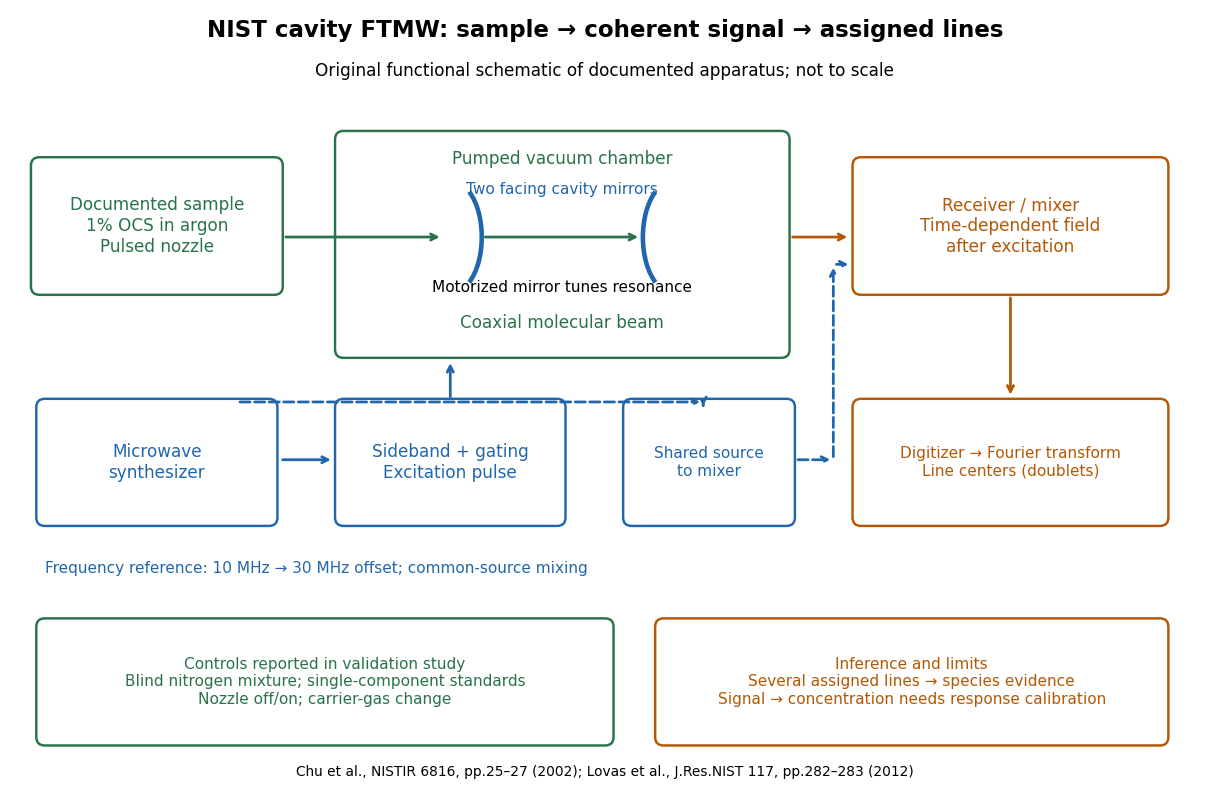
Original functional schematic of the NIST trace-gas FTMW validation experiment
. An original arrangement of the documented sample, cavity, excitation and readout functions; no published figure is copied. The frequency-reference branch summarizes the later portable electronics described in the NIST historical account. The sample fraction is a reported preparation, not a synthetic measured spectrum. Details.
The portable electronics used a common microwave source and a offset derived from a reference. Cavity tuning selects the accessible band; it does not independently calibrate the line's frequency. Absolute accuracy also requires a characterized reference and receiver offset.[ref]
Evolution of Microwave Spectroscopy at the National Bureau of Standards (NBS) and the National Institute of Standards and Technology (NIST) by F. J. Lovas and others. §5, printed pp. 282–283, Figure 20: source splitting, frequency reference, single-sideband modulation, mixer and digitizer
Reported controls included nozzle off/on, carrier-gas changes, and a blind nitrogen mixture. Assigned components were correctly identified. Single-component standards supplied preliminary concentration calibration; one standard per quantified component left response validation incomplete. Condensation and departure from thermal equilibrium in the nozzle were acknowledged limitations.[ref]
The Validation of Fourier-Transform Microwave Spectroscopy for Trace-Gas Analysis. printed pp. 26–27: nozzle/carrier comparison in Figure 6, blind mixture in Table II, one-standard calibration and nozzle-expansion limitations
The lesson is that rotational spectral line assignment and concentration inference answer different questions. Reproducible frequencies support molecular identity; converting signal size into concentration requires calibrated response under the same transfer and excitation conditions. The thermal absorption example above does not model a nozzle's nonequilibrium populations or the cavity's pulsed response.
A distinct experiment in An Automated, Highly Selective Reaction Monitoring Instrument Using Molecular Rotational Resonance Spectroscopy by Ann Adele Byars and others (Figure 1 and Figure 3) tracked exchange of hydrogen for deuterium in ethyl acetoacetate mixed with deuterated methanol. A pump circulated chemical solution through a valve; a calibrated loop injected aliquots into a temperature-controlled inlet, held at room temperature for this exchange experiment. Neon carried the vaporized sample to a cavity spectrometer. Separately recorded broadband spectra and fitted rotational Hamiltonians identified transitions for the four isotopic forms before targeted monitoring.[ref]
An Automated, Highly Selective Reaction Monitoring Instrument Using Molecular Rotational Resonance Spectroscopy by Ann Adele Byars and others. apparatus and assignment paragraphs around Figure 1; deuterium-exchange method and Figure 3
The readout was each assigned line's signal over reaction time, normalized by the sum of the four signals. Transitions with the same quantum numbers reduced response-factor differences. A solvent wash addressed observed sample carryover. The signals distinguished the unsubstituted, two singly substituted, and doubly substituted forms. Only one gas-phase structural form was detected; the authors explicitly caution that their readout does not recover the structural chemical equilibrium in chemical solution.[ref]
An Automated, Highly Selective Reaction Monitoring Instrument Using Molecular Rotational Resonance Spectroscopy by Ann Adele Byars and others. deuterium-exchange paragraphs surrounding Figure 3; sample carryover and wash paragraph
This example uses nonlinear molecules, so neither the single-constant linear rigid rotor nor the synthetic comb above is a fit to its spectrum. Its connection to the model is the sensitivity of moments of inertia to isotope placement. Line position helps assign a species; signal size additionally depends on sample transfer, initial quantum state populations and instrument response. Matching quantum numbers is a useful control, not a proof of identical response factors. These distinctions explain why resolving species and determining their concentrations are separate inference steps.
Articles by others on the same topic
There are currently no matching articles.